PGAM1-Chk1 interaction as a novel target for senolysis to preserve resilience during aging

Takumi Mikawa

Masahiro Kameda
Zhining Zhang
Hiroshi Kondoh
*
*Correspondence to:
Hiroshi Kondoh, Geriatric Unit, Graduate School of Medicine, Kyoto University, Kyoto 606-8501, Japan.
E-mail: hkondoh@kuhp.kyoto-u.ac.jp
Geromedicine. 2026;2:202625. 10.70401/Geromedicine.2026.0034
Received: April 09, 2026Accepted: August 17, 2026Published: August 17, 2026
Abstract
Aging is a complex biological process, which is affected by several factors, lifestyle, diet, genetic or epigenetic factors, environmental stress, and metabolisms. Among others, several reports observed increased glycolysis in senescent cells (SnCs) and individual aging, whose causal effects or biological mechanisms have been unclear. Recently, we identified phosphoglycerate mutase 1 (PGAM1)-checkpoint kinase 1 (Chk1) binding as a booster for glycolytic metabolism and cell viability in SnCs. Secretory phenotype of inflammatory factors, known as senescence associated secretory phenotype (SASP), is one of prominent properties in SnCs, which accelerates chronic inflammation and aging-relevant dysfunctions in tissues. Inhibition of PGAM1-Chk1 binding removes SnCs and suppresses SASP, alleviating organ damage and pulmonary fibrosis in vivo. Thus, PGAM1-Chk1 interaction represents a target for senolysis to preserve resilience in aging.
Keywords
Resilience, senescence associated secretory phenotype, senolysis, glycolysis, PGAM1, Chk1, HIF-2α, FoxM1
1. Introduction
Many organisms maintain normal tissue functions and physiological homeostasis under various environmental changes and stresses, by the spare ability of adaptation, response and recovery, called “resilience”[1,2]. In older adults or frailty patients, who are vulnerable to stress and bed-ridden states, the resilience is altered and impaired, leading to functional decline of organs and development of age-related diseases. It has been assumed that the accumulating senescent cells (SnCs) in aged tissues largely affect in vivo resilience[3,4]. SnCs display several hallmarks of cellular senescence[5,6]; positivity of senescence-associated β-galactosidase (SA-β-GAL staining)[7,8], widespread cytoplasm with large nucleus[9-11], DNA damage including telomere erosion[12-14], mitochondrial dysfunction[15,16], impaired autophagy[17], senescence-associated heterochromatin foci (SAHF)[18] and so on. In addition, due to the activated programs of transcription factor nuclear factor (NF-κB), SnCs acquire enhanced anti-apoptotic potency, compared to that of young cells, followed by enhanced inflammatory programs of NF-κB as senescence-associated secretory phenotype (SASP) (Figure 1)[19-21]. Thus, chronic inflammation stemming from SnCs is supposed to cause a decrease in the resilience of aged individuals[22].
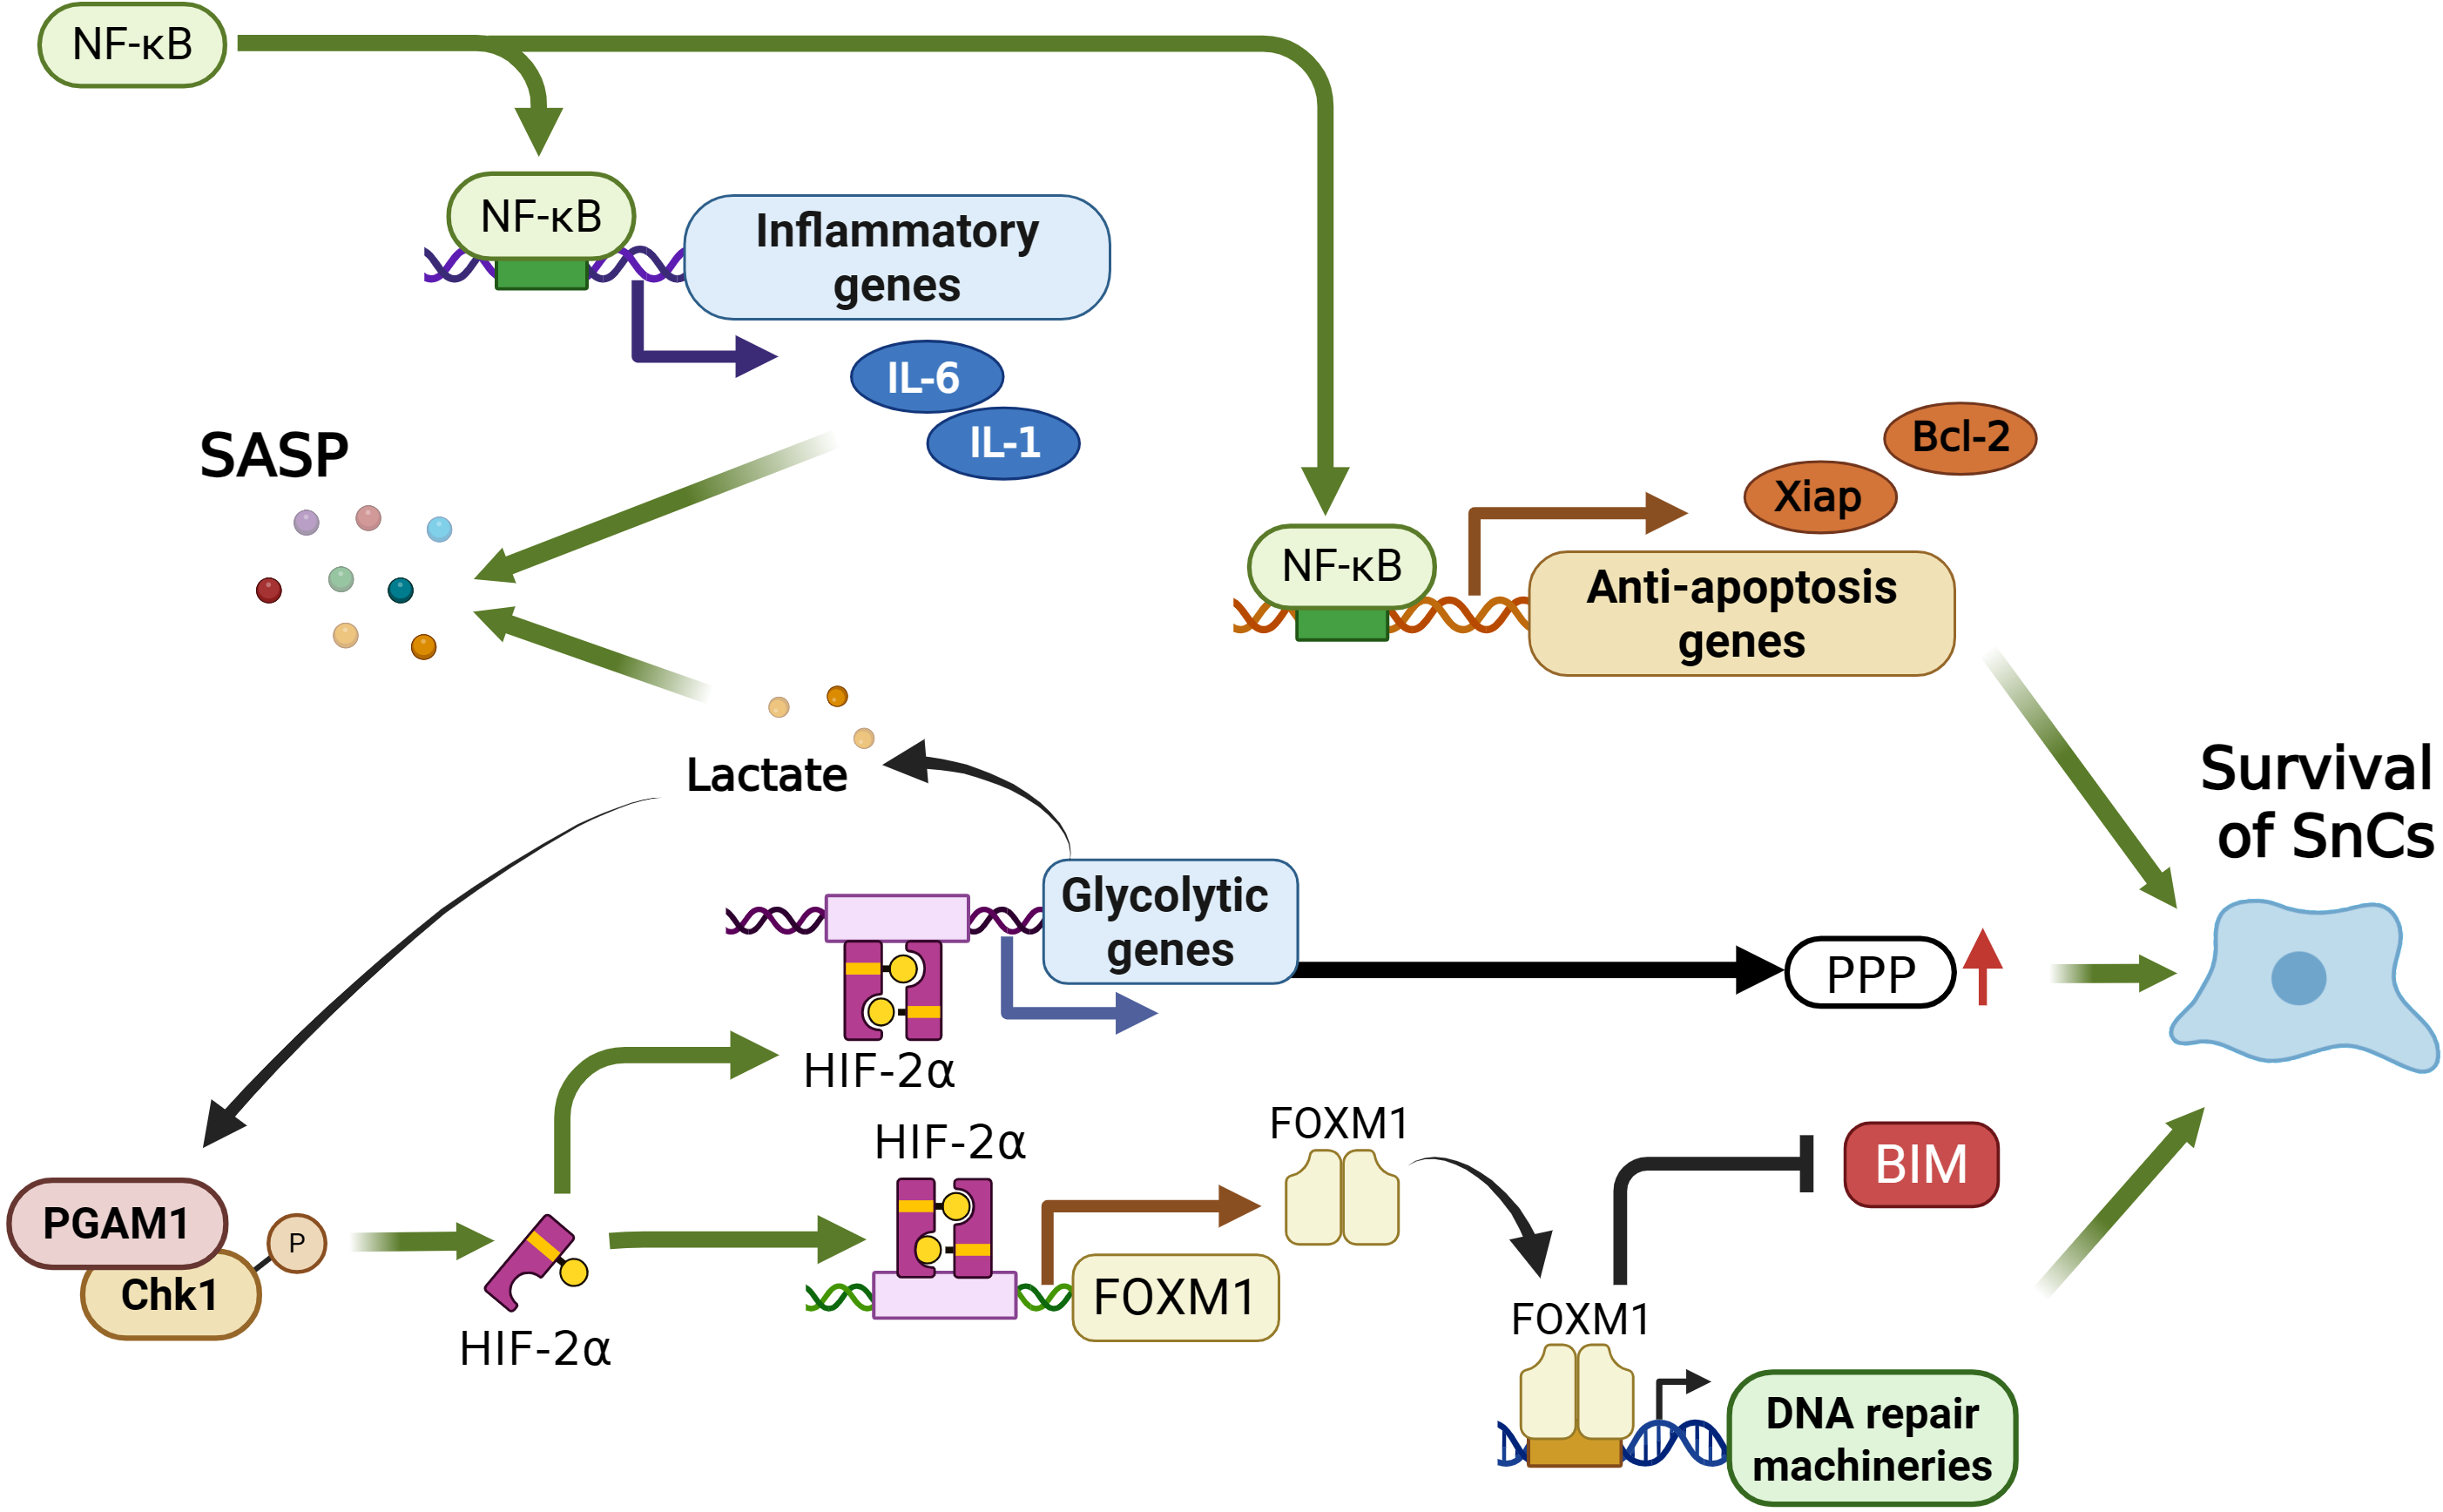
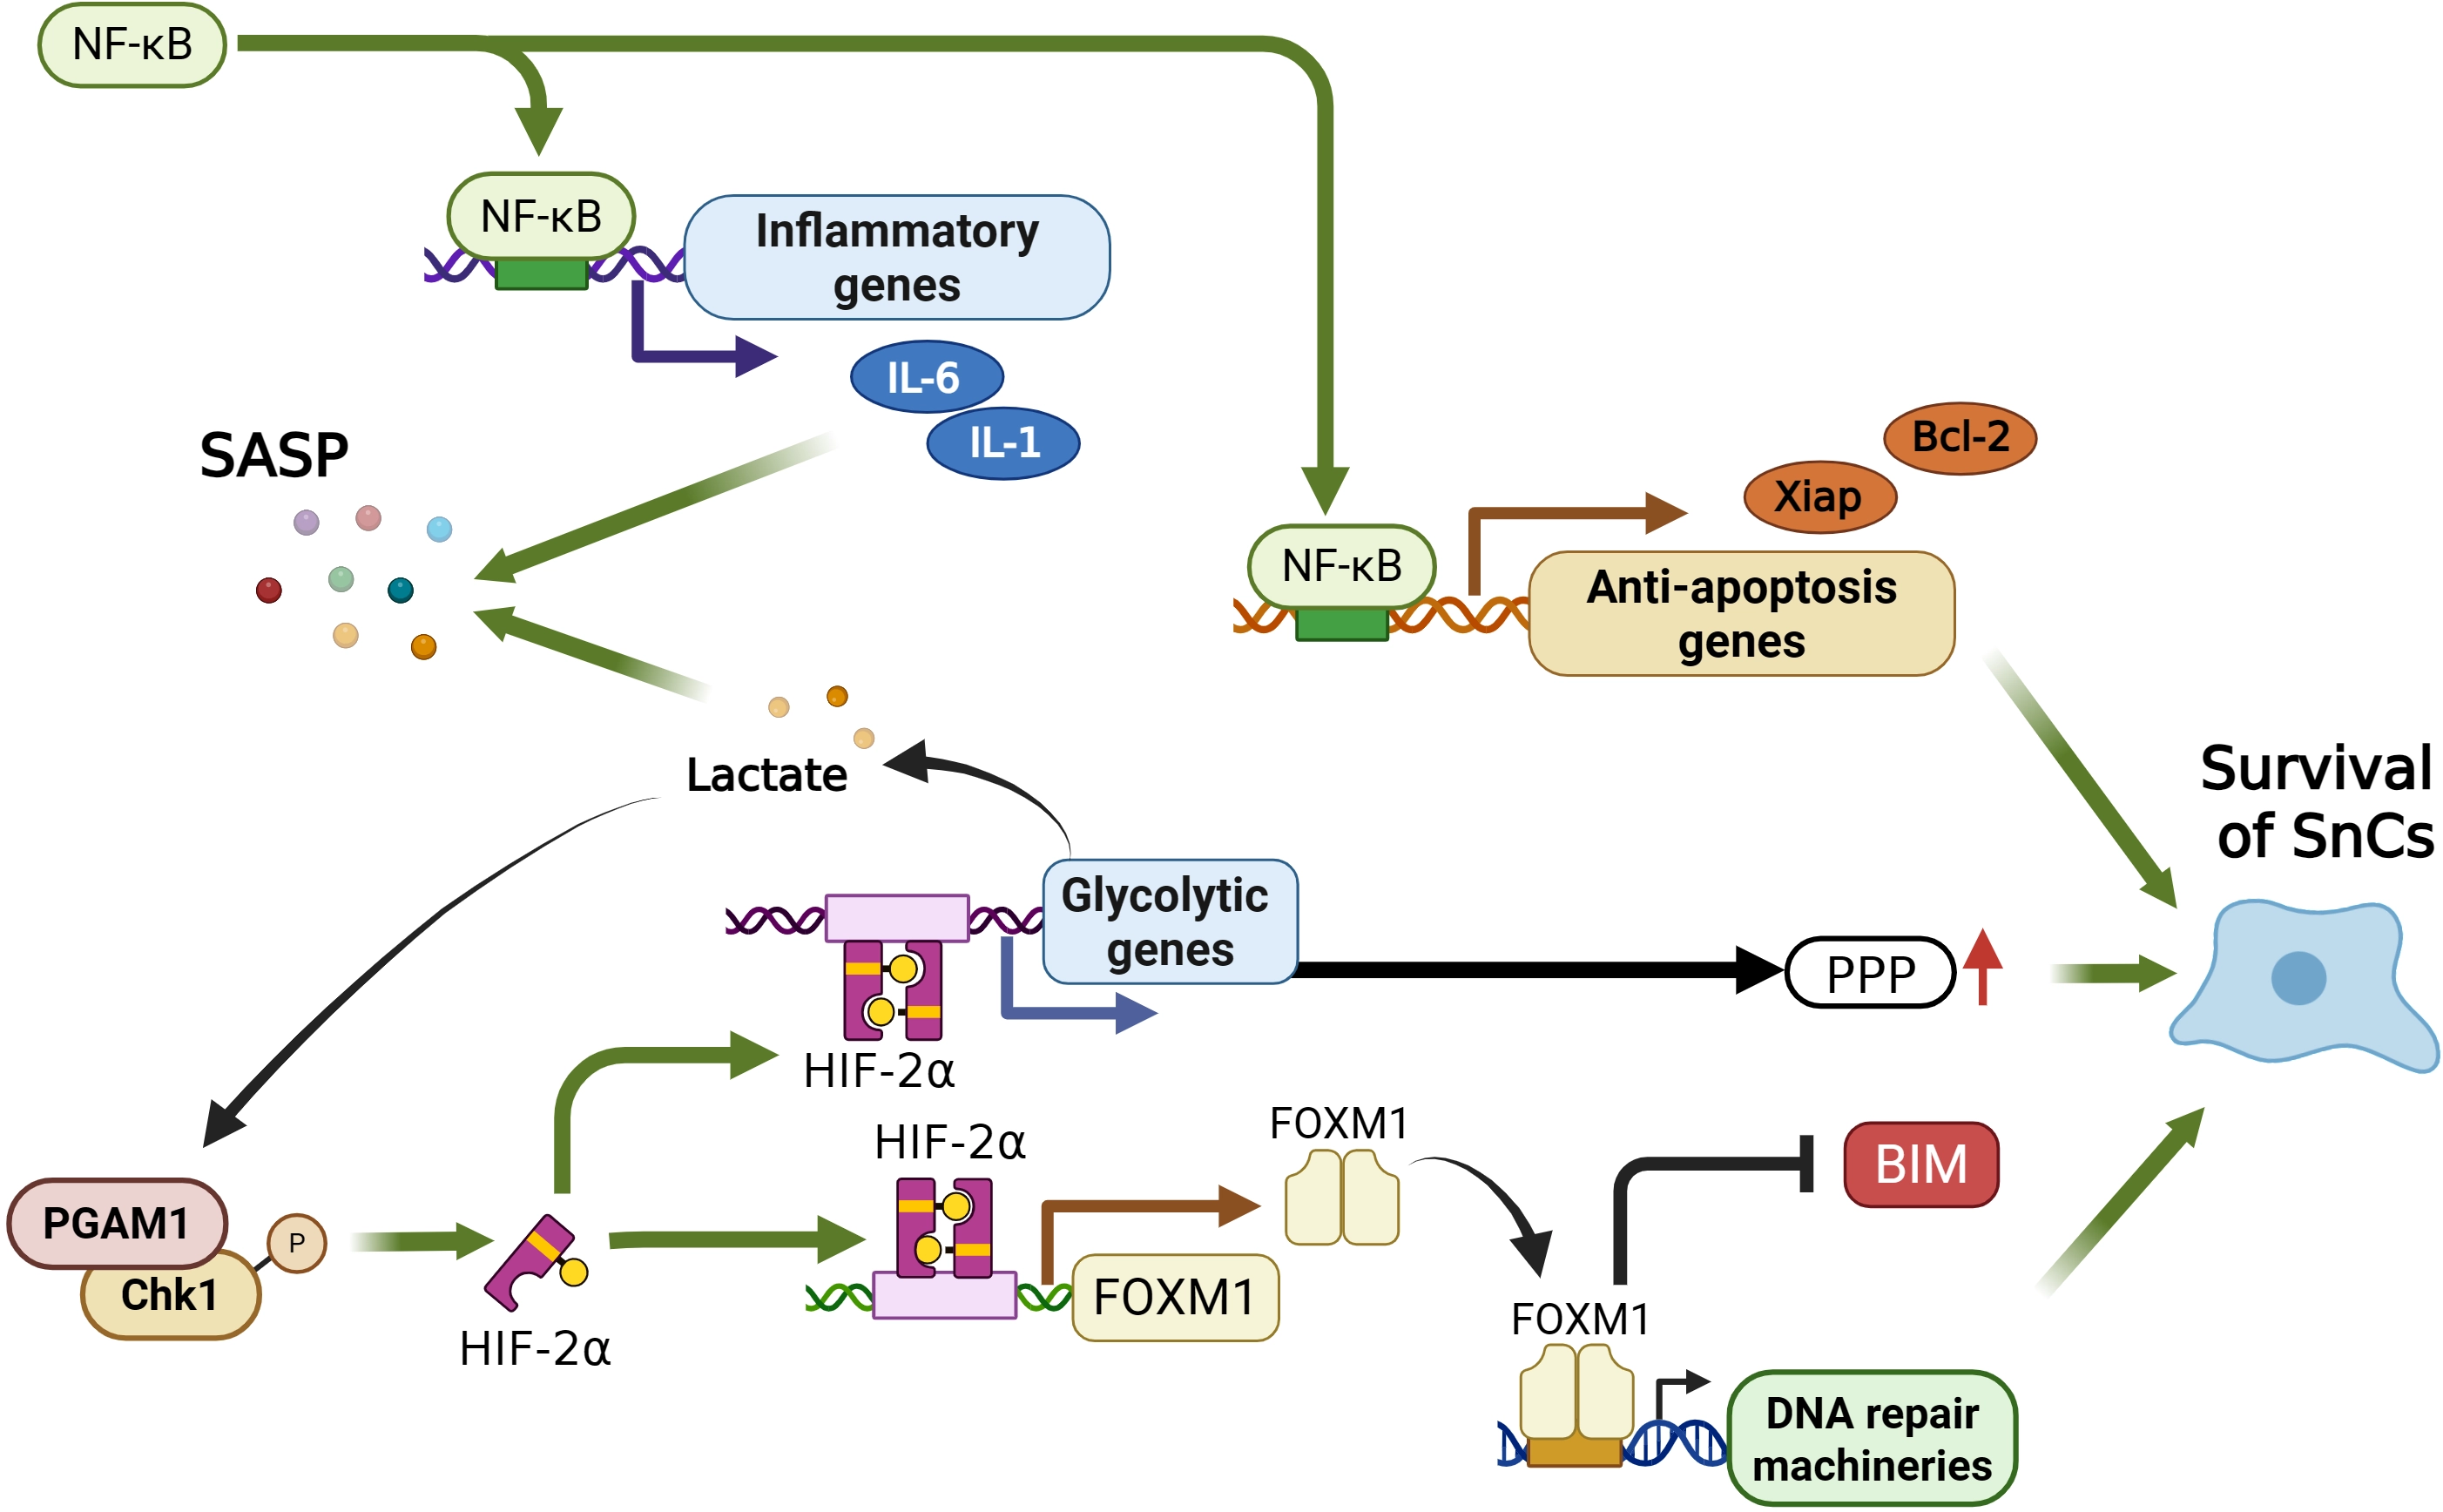
Figure 1. Transcriptional program for survival and SASP of SnCs. Transcriptional NF-κB is operating in SnCs, resulting in activation of anti-apoptotic genes (e.g. Bcl-2 and Xiap) and a secretory phenotype, SASP. Senescence-inducing stress also enhances PGAM1-Chk1 binding, which stabilizes and activates HIF-2α. The pseudo-Warburg effect in SnCs is coupled with enhanced PPP, providing nucleotide pools, and secretion of lactate as an enhancer of PGAM1-Chk1 binding. HIF-2α also activates FoxM1 in SnCs, which suppresses proapoptotic BIM and activates DNA repair machineries. Created in BioRender. Kondoh, H. (2026) https://app.biorender.com/illustrations/6792f262706c9c5c43d6cc8c. SASP: senescence associated secretory phenotype; SnCs: senescent cells; NF: nuclear factor; Bcl-2: B-cell lymphoma 2; Xiap: X-linked inhibitor of apoptosis protein; PGAM1: phosphoglycerate mutase 1; Chk1: checkpoint kinase 1; HIF: hypoxia-inducible factor; PPP: pentose phosphate pathway; FoxM1: Forkhead box M1; IL-6: interleukin-6.
To restore resilience during the aging process, a new approach called “senotherapy”, targeting the properties of aging for therapy, was proposed[23,24]. Among others, selective removal of SnCs is defined as “senolysis”. For example, an inhibitor of the anti-apoptotic gene B-cell lymphoma 2 (Bcl-2), ABT263, has been reported as a senolytic drug[25,26]. It is crucial to elucidate how senolytic effects are mechanistically coupled with properties of SnCs. Moreover, as “heterogeneity” of SnCs in vivo is noted[27], detailed verification for selectivity of senolysis against subtype of SnCs is needed to apply it to human clinical practice with minimization of the side effects.
2. Biological Significance of Increased Glycolytic Metabolism in SnCs
Abnormal and impaired mitochondrial function was well known in aged individuals and cellular aging, while much remains unknown about glycolytic metabolism. Although several reports observed increased glycolysis in SnCs and aged organisms[28-31], it has been unclear why and how glycolysis is biologically connected to senescent phenotype.
Enhanced glycolysis in localized lesions is well observed in many cancers, called the Warburg effect[32,33]. Previously, we found that in cancer cells with oncogenic Ras activation, the Warburg effect was maintained by “non-enzymatic function” of glycolytic enzyme phosphoglycerate mutase (PGAM), that is, increased PGAM1-checkpoint kinase 1 (Chk1) binding[34]. Applying NanoLuc Binary Technology (NanoBiT), a two-subunit system based on NanoLuc luciferase that can be used for intracellular detection of protein-protein interactions (PPIs)[35,36], enabled us to effectively evaluate PGAM1-Chk1 binding. We noticed that oncogenic Ras-induced SnCs also displayed both increased PGAM1-Chk1 binding and enhanced glycolysis, designated as the pseudo-Warburg effect[37]. It has been conceived that the Warburg effect in cancers meets the increasing energy demand of rapidly-proliferating cells. But, as SnCs are suffering permanent cell cycle arrest, the pseudo-Warburg effect in SnCs would be provoked by other mechanisms. Alternatively, it might be possible that the pseudo-Warburg effect compensates for the mitochondrial dysfunction, observed in SnCs. However, the inhibition of the mitochondrial electron transport chain with rotenone or antimycin A failed to induce PGAM1-Chk1 binding in primary cells, suggesting that PGAM1-Chk1 binding seems not to be coupled with energy demand or mitochondria status[37]. Importantly, the pseudo-Warburg effect was crucial for cellular survival not only in oncogene-induced senescence, but also in DNA damage-induced, irradiation-induced, and replicative senescence[37]. Thus, senescence-inducing stress provokes the pseudo-Warburg effect in SnCs, while the classical Warburg effect meets the requirements of greedy cancerous cells. Collectively, we identified enhanced PGAM1-Chk1 interaction as one of prominent features of SnCs.
Instead, we identified transcription factor hypoxia-inducible factor (HIF)-2α as a target of PGAM1-Chk1 binding. In agreement with the activation of HIF-2α in SnCs, glycolytic pathway is upregulated, associated with activation of the pentose phosphate pathway (PPP) and production of lactate. PPP plays critical roles in supplying nucleotide precursors, thereby supporting DNA integrity[38], while lactate is well established as clinical biomarker of hypoxia, e.g. ischemic stroke[39]. Lactate enhances PGAM1-Chk1 binding through signaling of its G protein-coupled receptor 81 (GPR81) receptor. Interestingly, recent study suggested that HIF-2α accelerates SASP as a priming factor[40]. Thus, HIF-2α can upregulate both inflammatory SASP factors and lactate in parallel. These findings suggest PGAM1-Chk1 mediated glycolysis in SnCs is biologically crucial to maintain both DNA integrity and its binding, via PPP and lactate, respectively (Figure 1).
3. Senolytic Mechanism by Inhibition of PGAM1-Chk1 Binding
Nutlin was developed as an anticancer agent that inhibits tumor protein p53 (p53)-mouse double minute 2 (MDM2) binding[41]. We had previously reported ubiquitination of PGAM1 mediated by Mdm2[42]. Initially, we suspected the potential of Nutlin as an inhibitor of the binding between PGAM1 and Mdm2 ligase, but it proved to be completely ineffective. Based on the information on the Nutlin target sequence reported in the proteomics study by Nicholson et al.[43], we noticed two repeats of the Nutlin target sequence at the N-terminus of the PGAM1, which Chk1 binds. In fact, we have identified Nutlin as an inhibitor for PGAM1-Chk1 binding[34]. Although Nutlin was structurally designed to inhibit the p53-binding pocket, little was known about the structural basis of PGAM1-Chk1 binding, which remains to be resolved.
Interestingly, while Nutlin composes two optical isomers (3a and 3b), the efficiency for inhibition of p53-MDM2 binding differs more than 100-fold between Nutlin 3a and 3b[41]. On the other hand, 3a and 3b can inhibit PGAM1-Chk1 binding to the same extent. Nutlin 3a, activator of p53, has high anti-cancer activity, and is highly toxic in normal cells and young mice in our study. However, Nutlin 3b selectively removed SnCs, but not young cells, and its safety in young mice was confirmed (Figure 2). These functional differences of two optical isomers, like Nutlin-3a and -3b described here, are not exceptional and are also reported in several other cases; e.g. ofloxacin vs levofloxacin as antibiotics[44]. These findings suggest Nutlin 3b, not 3a, as a potential senolytic drug. Indeed, Nutlin 3b alleviated several aging-relevant disorders in aged mice, muscle weakening and disorder in aged liver, kidney, and lung.
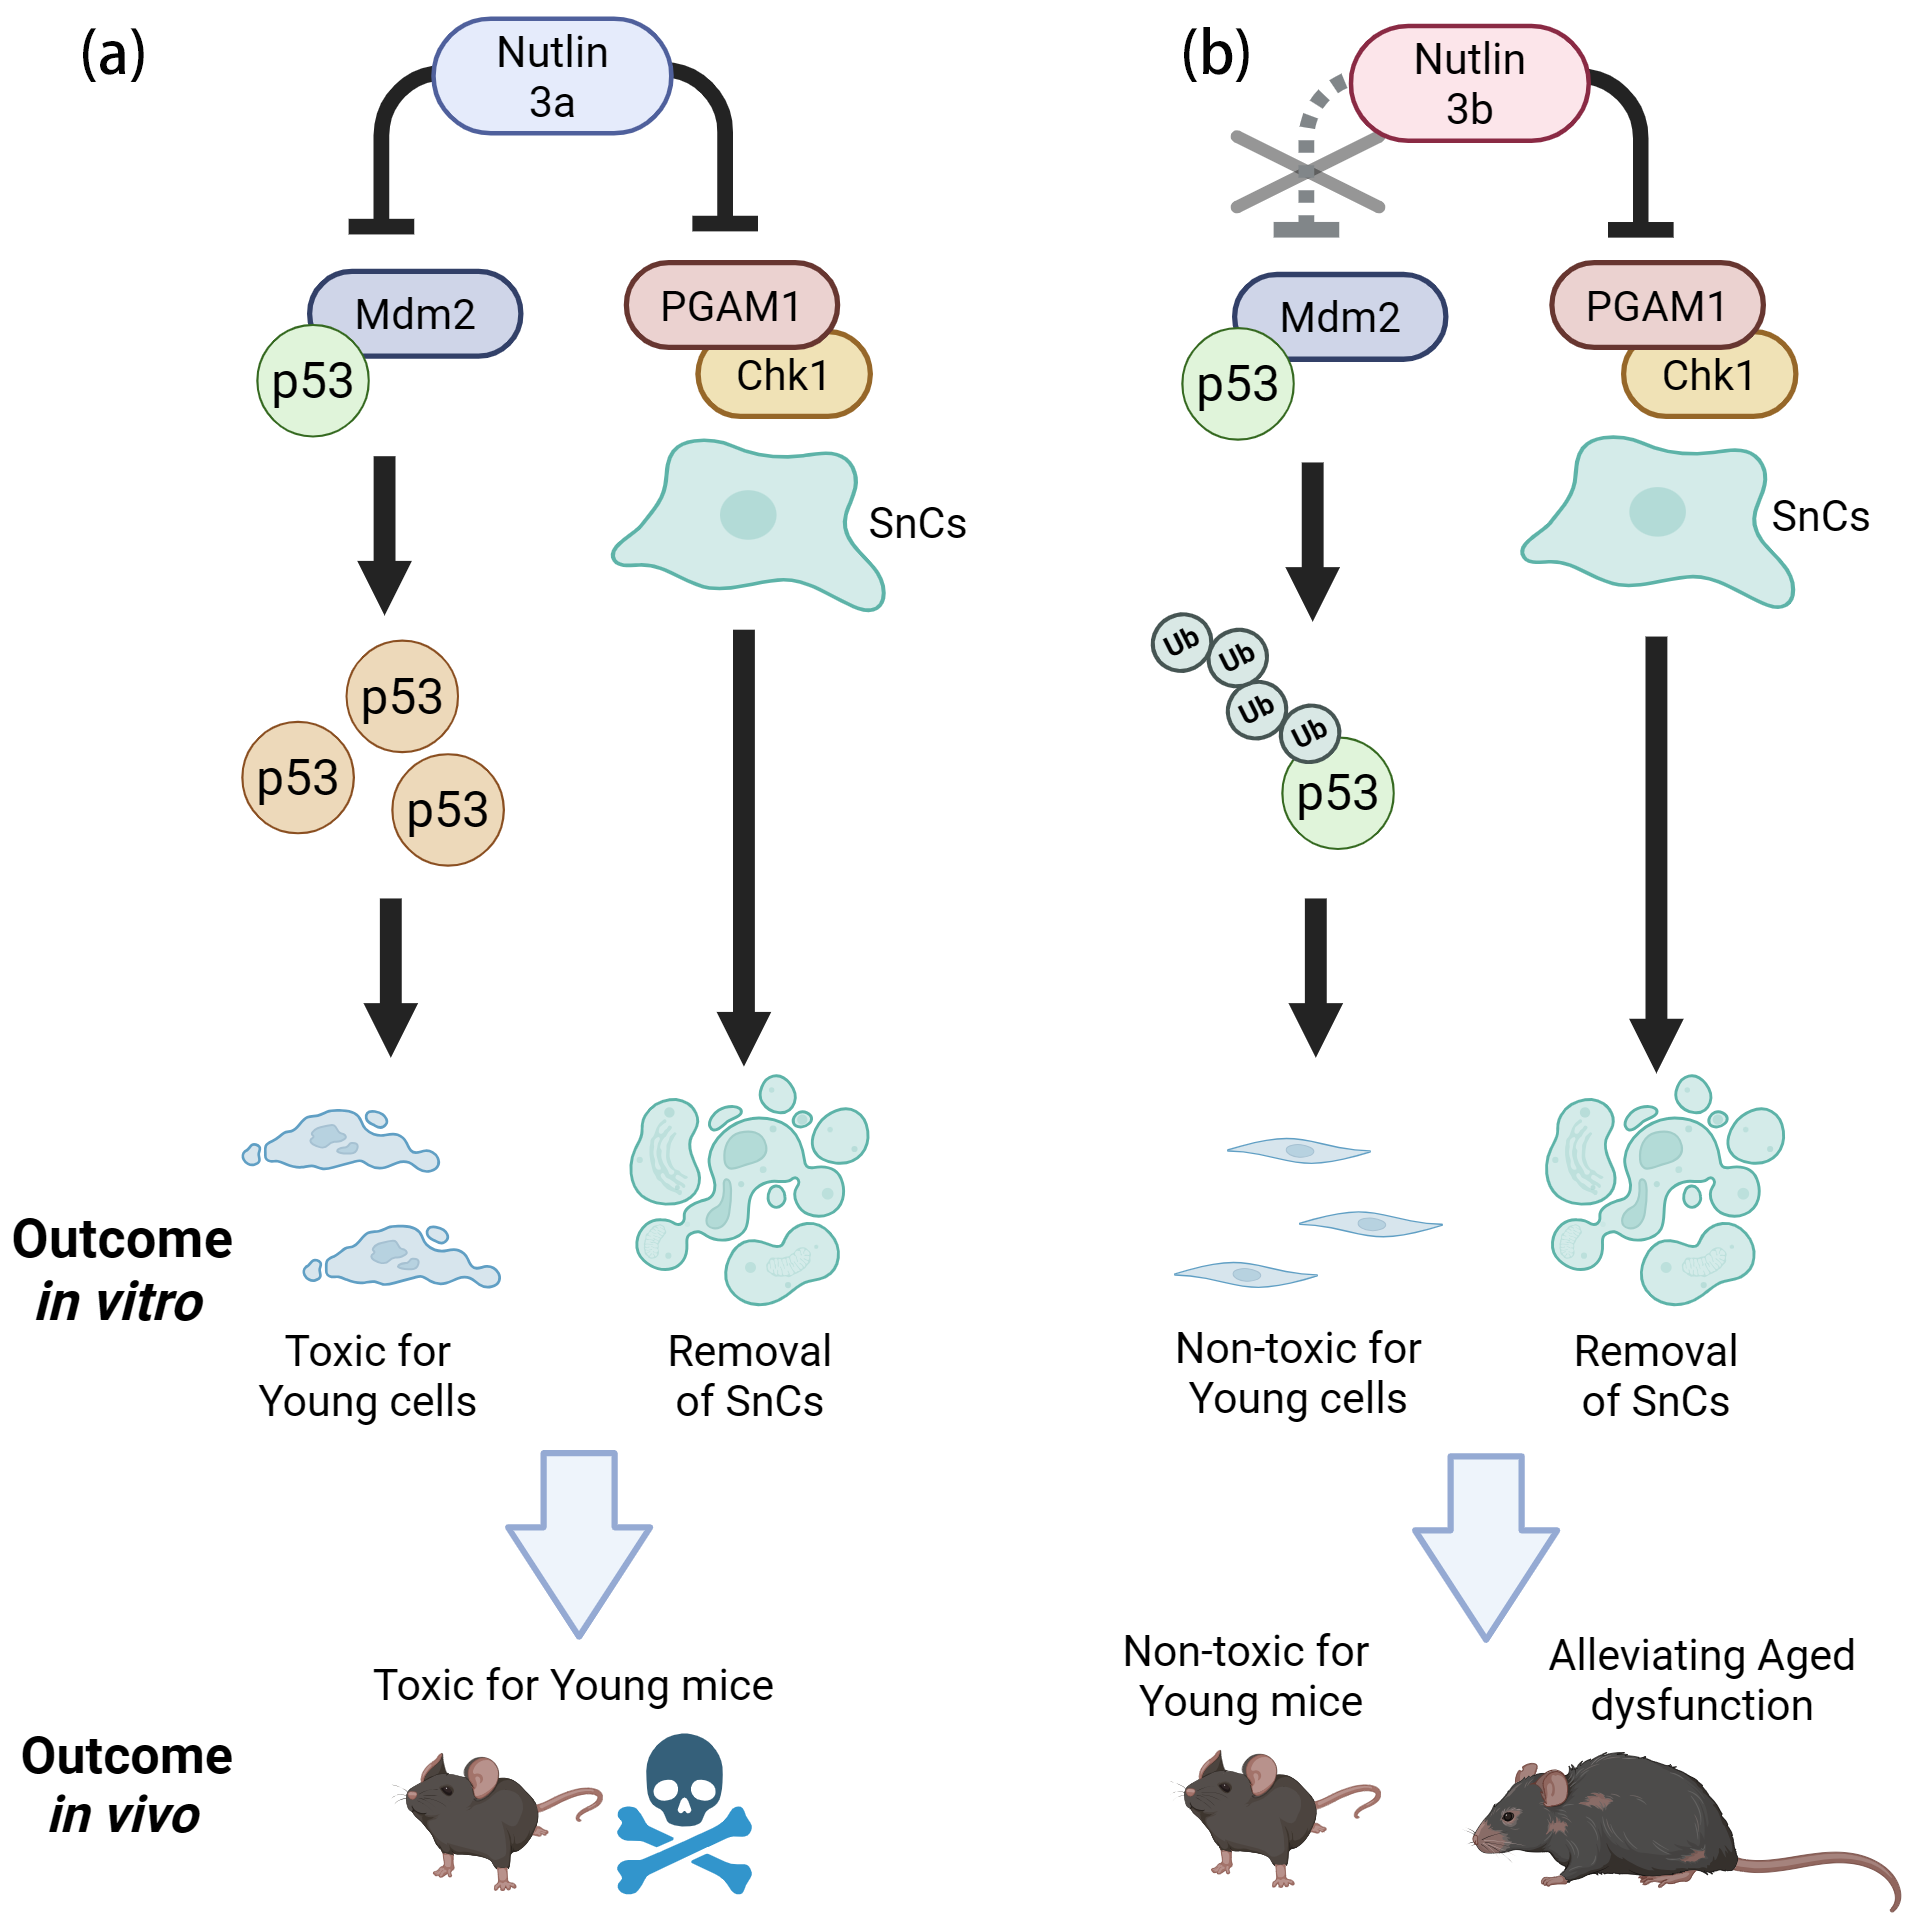
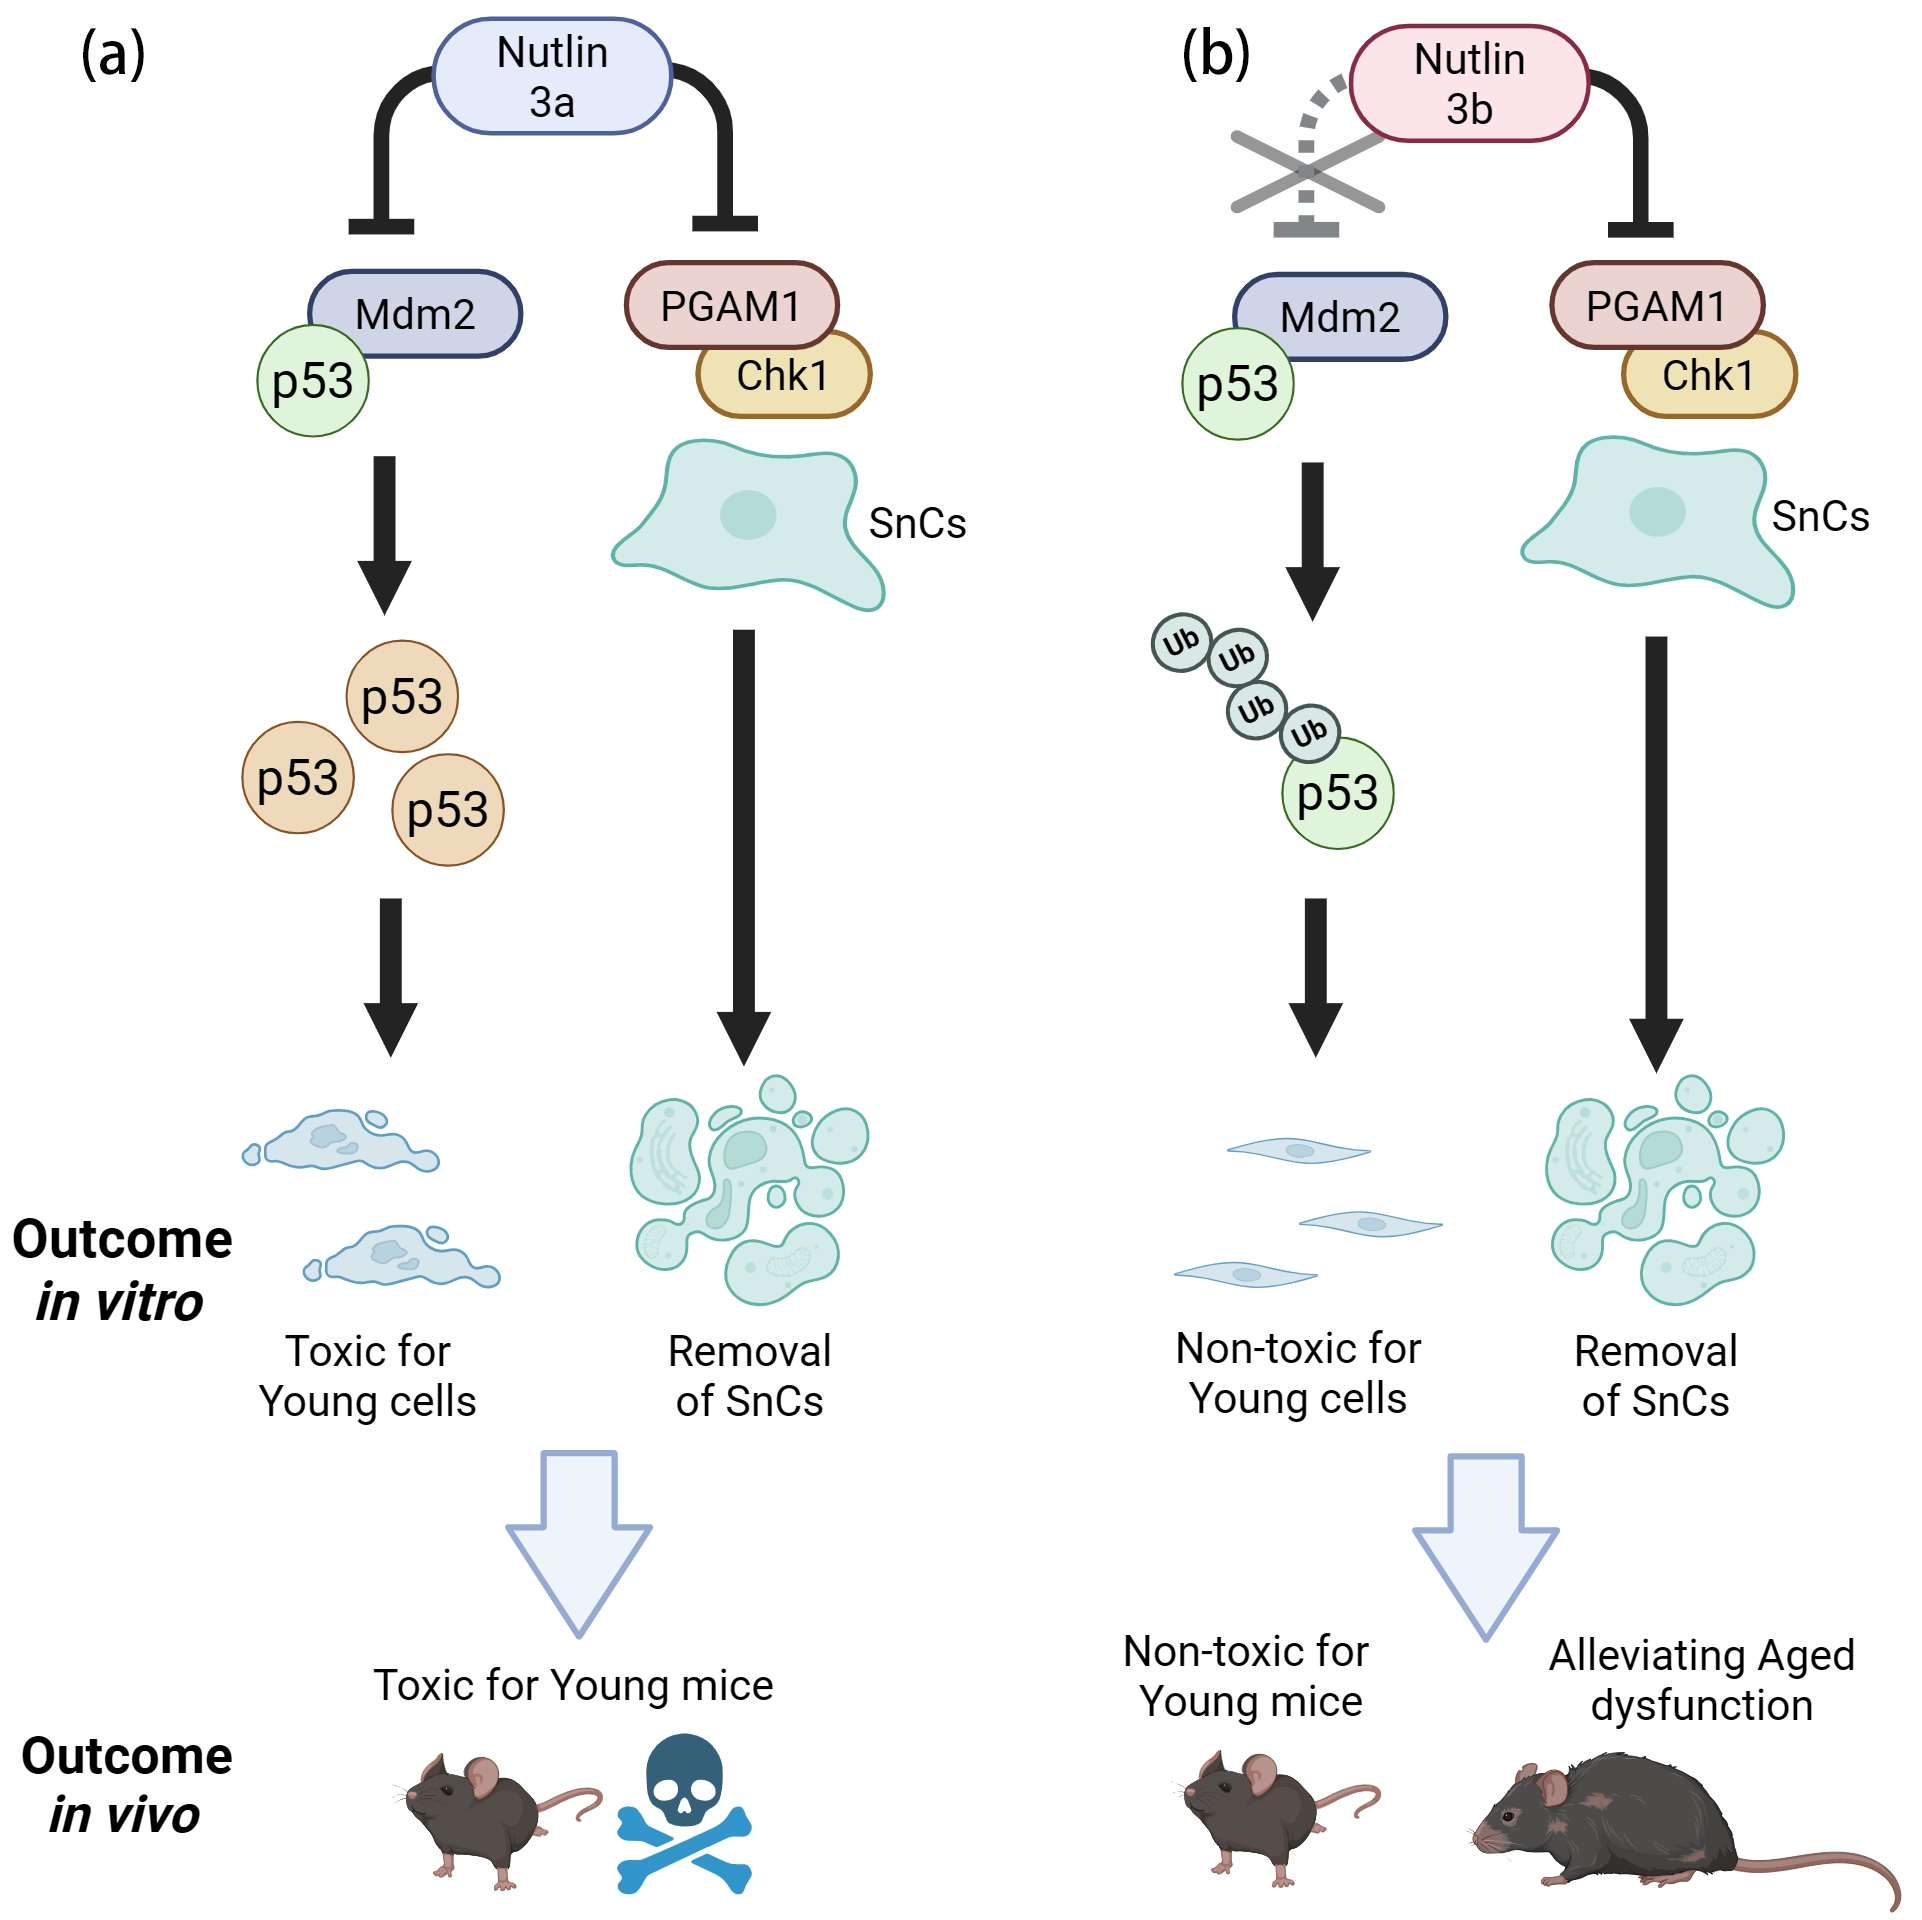
Figure 2. Comparison of biological effects between Nutlin 3a and 3b. (a) Nutlin 3a efficiently inhibits the interactions of both p53-Mdm2 and PGAM1-Chk1. Consequently, Nutlin 3a is rather toxic for young cells and mice, due to activation of p53; (b) Nutlin 3b interferes with PGAM1-Chk1 binding, not with that of p53-Mdm2. Nutlin 3b is effective for senotherapy in aged mice, but not toxic for young mice. Created in BioRender. Kondoh, H. (2026) https://app.biorender.com/illustrations/69bea4ad76a6c6dd199a49b8. p53: tumor protein p53; Mdm2: mouse double minute 2; PGAM1: phosphoglycerate mutase 1; Chk1: checkpoint kinase 1; SnCs: senescent cells; Ub: ubiquitin.
Cellular fates for survival are largely based on the balance between the pro-apoptotic and anti-apoptotic factors. Frontier works (e.g. Chang et al.[25]) reported that several “anti-apoptotic defenses” (e.g. Bcl-2 family) are actively operating in SnCs. These works inspired the challenges of senolysis, leading to identification of ABT263 or Dasatinib plus Quercetin (D + Q)[24]. Since then, several classes of senolysis drugs were identified (Figure 3)[45,46]. Inhibitors against BCL family proteins (BCL-XL, BCL-2, and BCL-W) comprise ABT analogues (ABT263 and ABT737), Nav-Gal, PZ15227, and Gingerenone A, while activators for p53 (FOXO-DRI, UBLX0101, P5091, and P22077) and inhibitors of proteostasis proteins, including heat shock protein 90 (HSP90) or coat protein complex I (COPI), 17-DMAG, IMP1088, and PCLX-001, are also effective for senolysis. In addition, senescence-specific membranous proteins, urokinase plasminogen activator receptor (uPAR) or glycoprotein nonmetastatic melanoma protein B (GPNMB), upstream signaling modules for survival (receptor tyrosine kinase (RTK) or phosphoinositide 3-kinase/protein kinase (BPI3K/AKT)), and metabolic enzyme glutaminase 1 (GLS1) are efficient targets of senolysis. On the other hand, the senolytic effect of Nutlin 3b is partly due to the activation of the proapoptotic gene Bim, but not of p53, while ABT263 inhibits Bcl2, an anti-apoptotic gene. The comparison between the transcriptome of Nutlin 3b-treated SnCs and more than 20,000 chromatin immunoprecipitation (ChIP) datasets registered in ChIP Atlas[47] (https://chip-atlas.org/) identified Forkhead box M1 (FoxM1) as the transcription factor with highest scores of fold enrichment (FE). Interestingly, many DNA repair factors are the target of FoxM1 in SnCs, in addition to known cell cycle factors. Thus, PGAM1-Chk1 axis suppresses “proapoptotic factors” BIM and upregulates DNA repair machineries by transcriptional FoxM1; notably, PCC1 or cardiac glycoside activates the other proapoptotic Puma or Noxa. As HIF-2α activates FoxM1 in SnCs, HIF-2α supports DNA integrity by activating both PPP and DNA repair machineries (Figure 1), while epigenetic factor bromodomain-containing protein 4 (BRD4), a target of senolytic ARV825, also regulates DNA repair machinery in SnCs[48]. It is possible that the enhanced “pro-apoptotic responses”, e.g. by Nutlin 3b, might synergistically function with suppression of “anti-apoptotic factors”, e.g. by ABT263 or D + Q, for induction of senolysis.
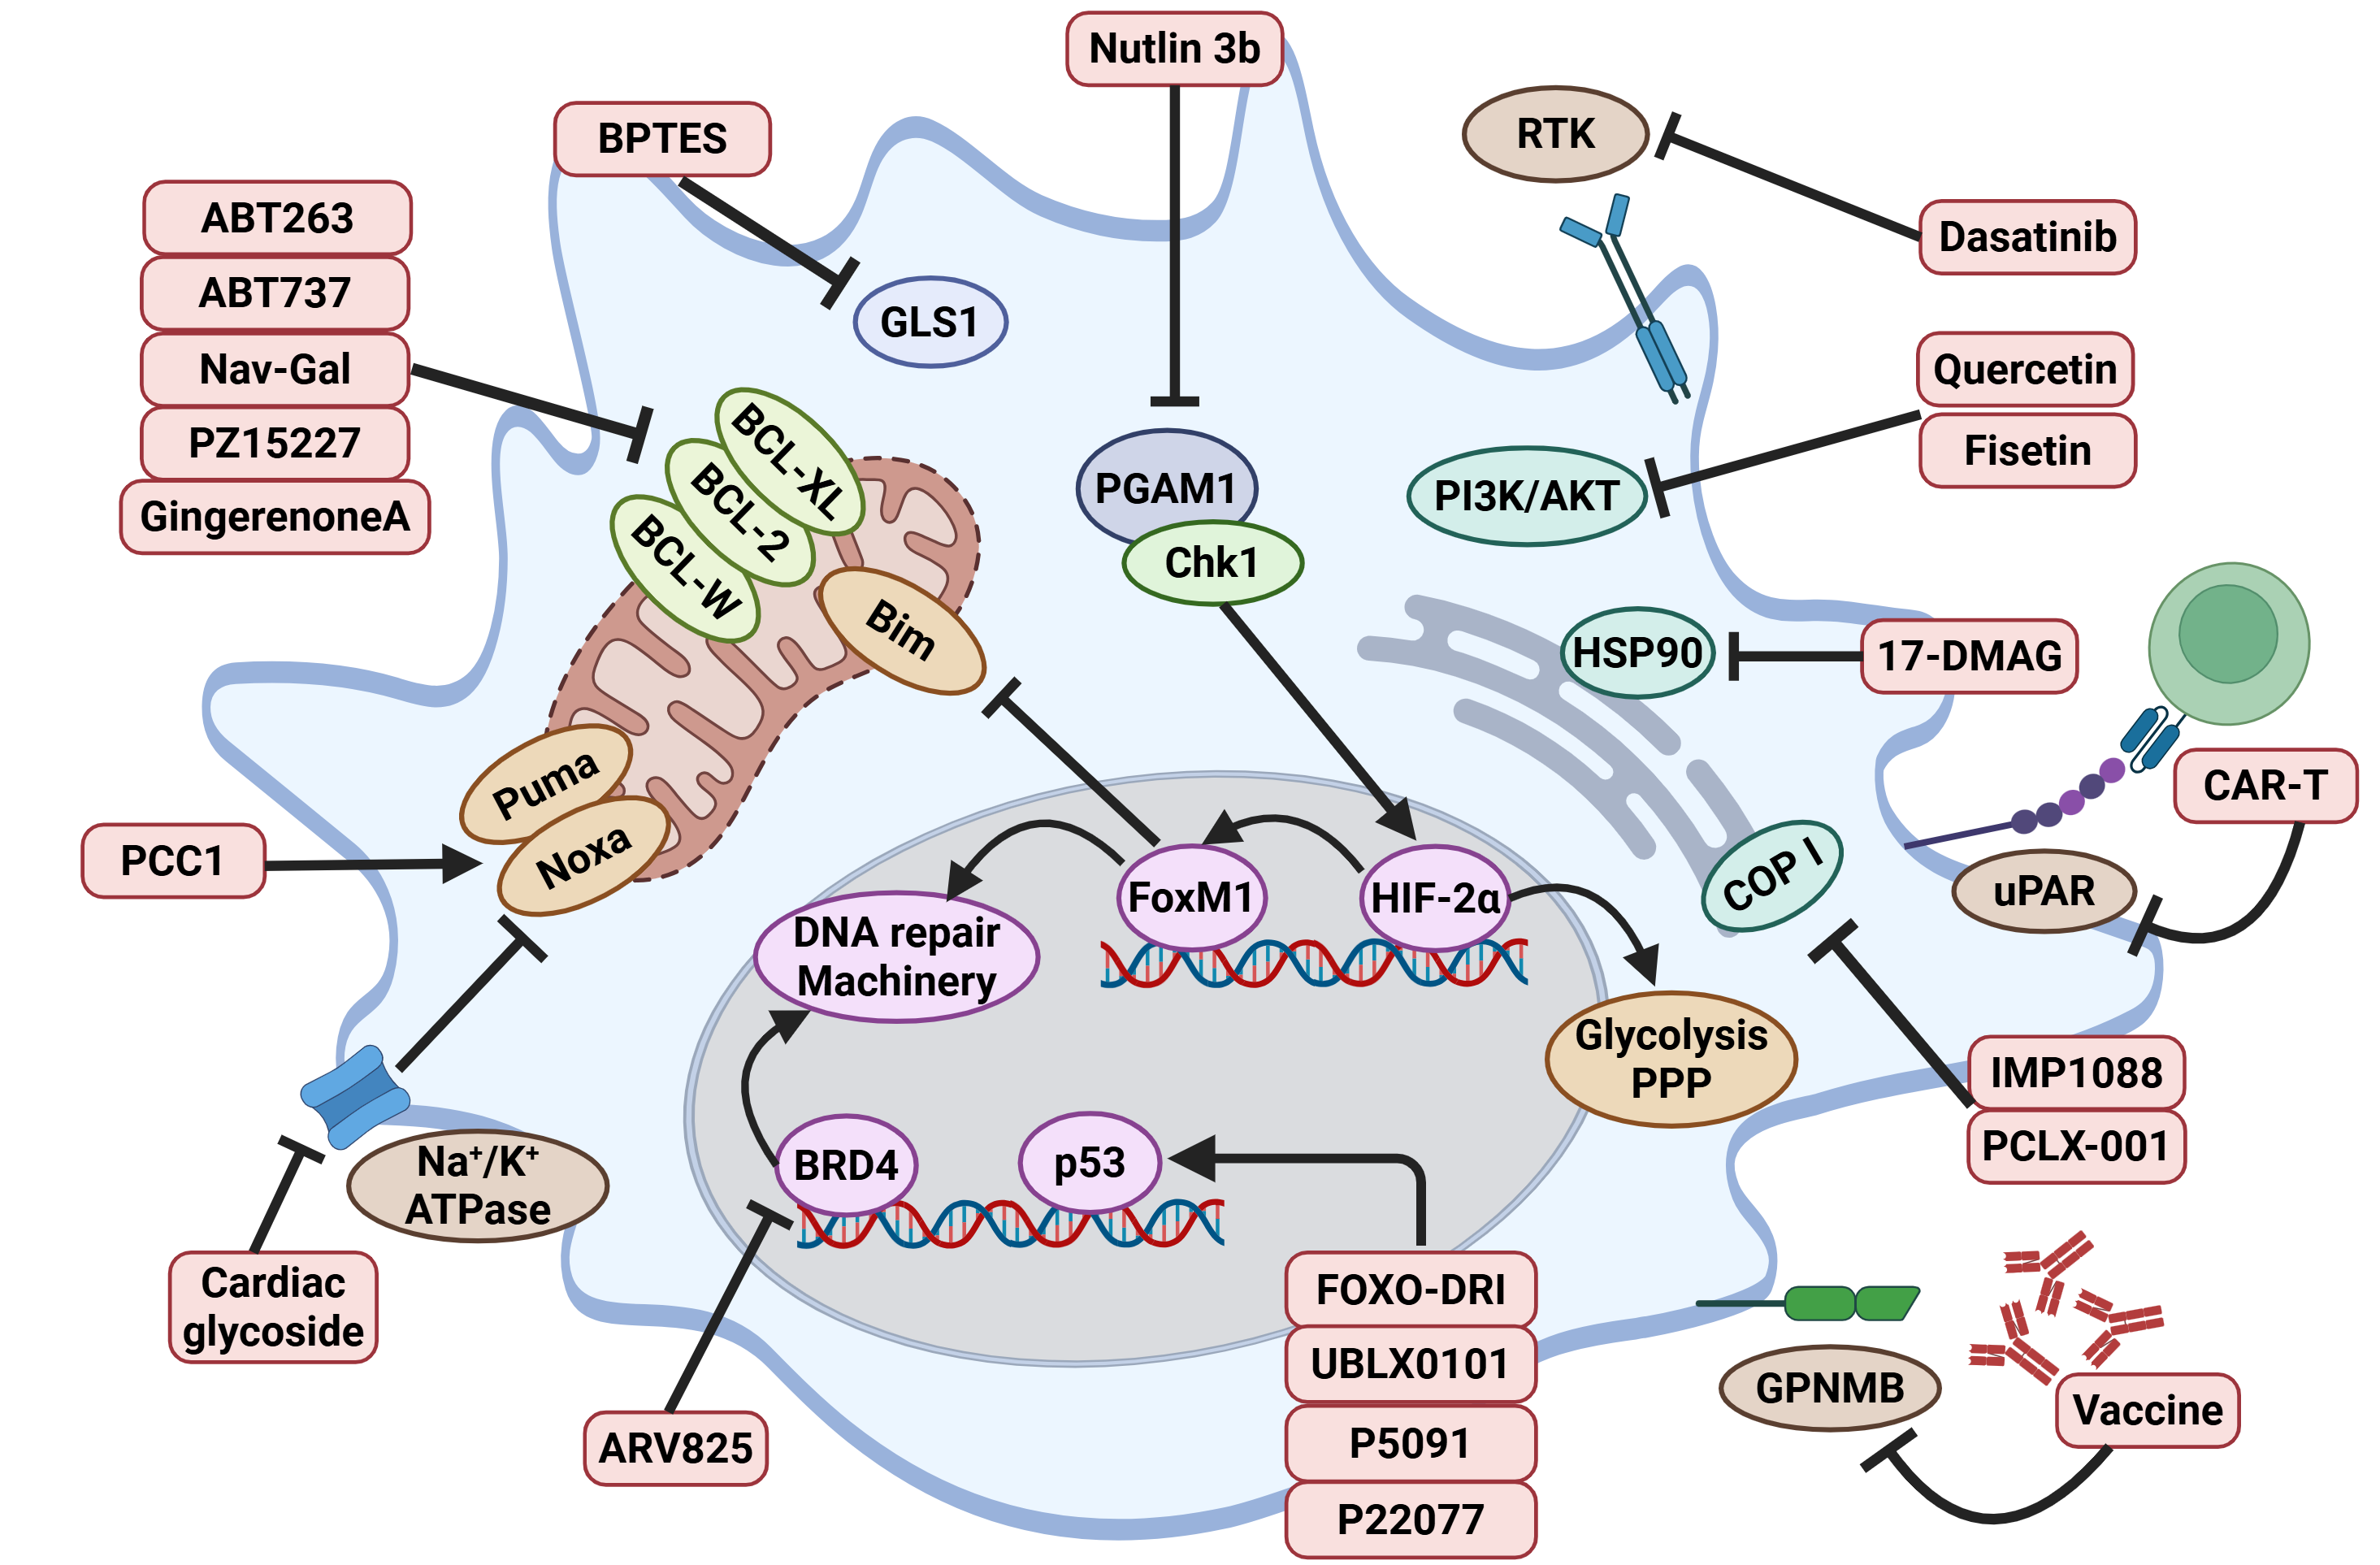
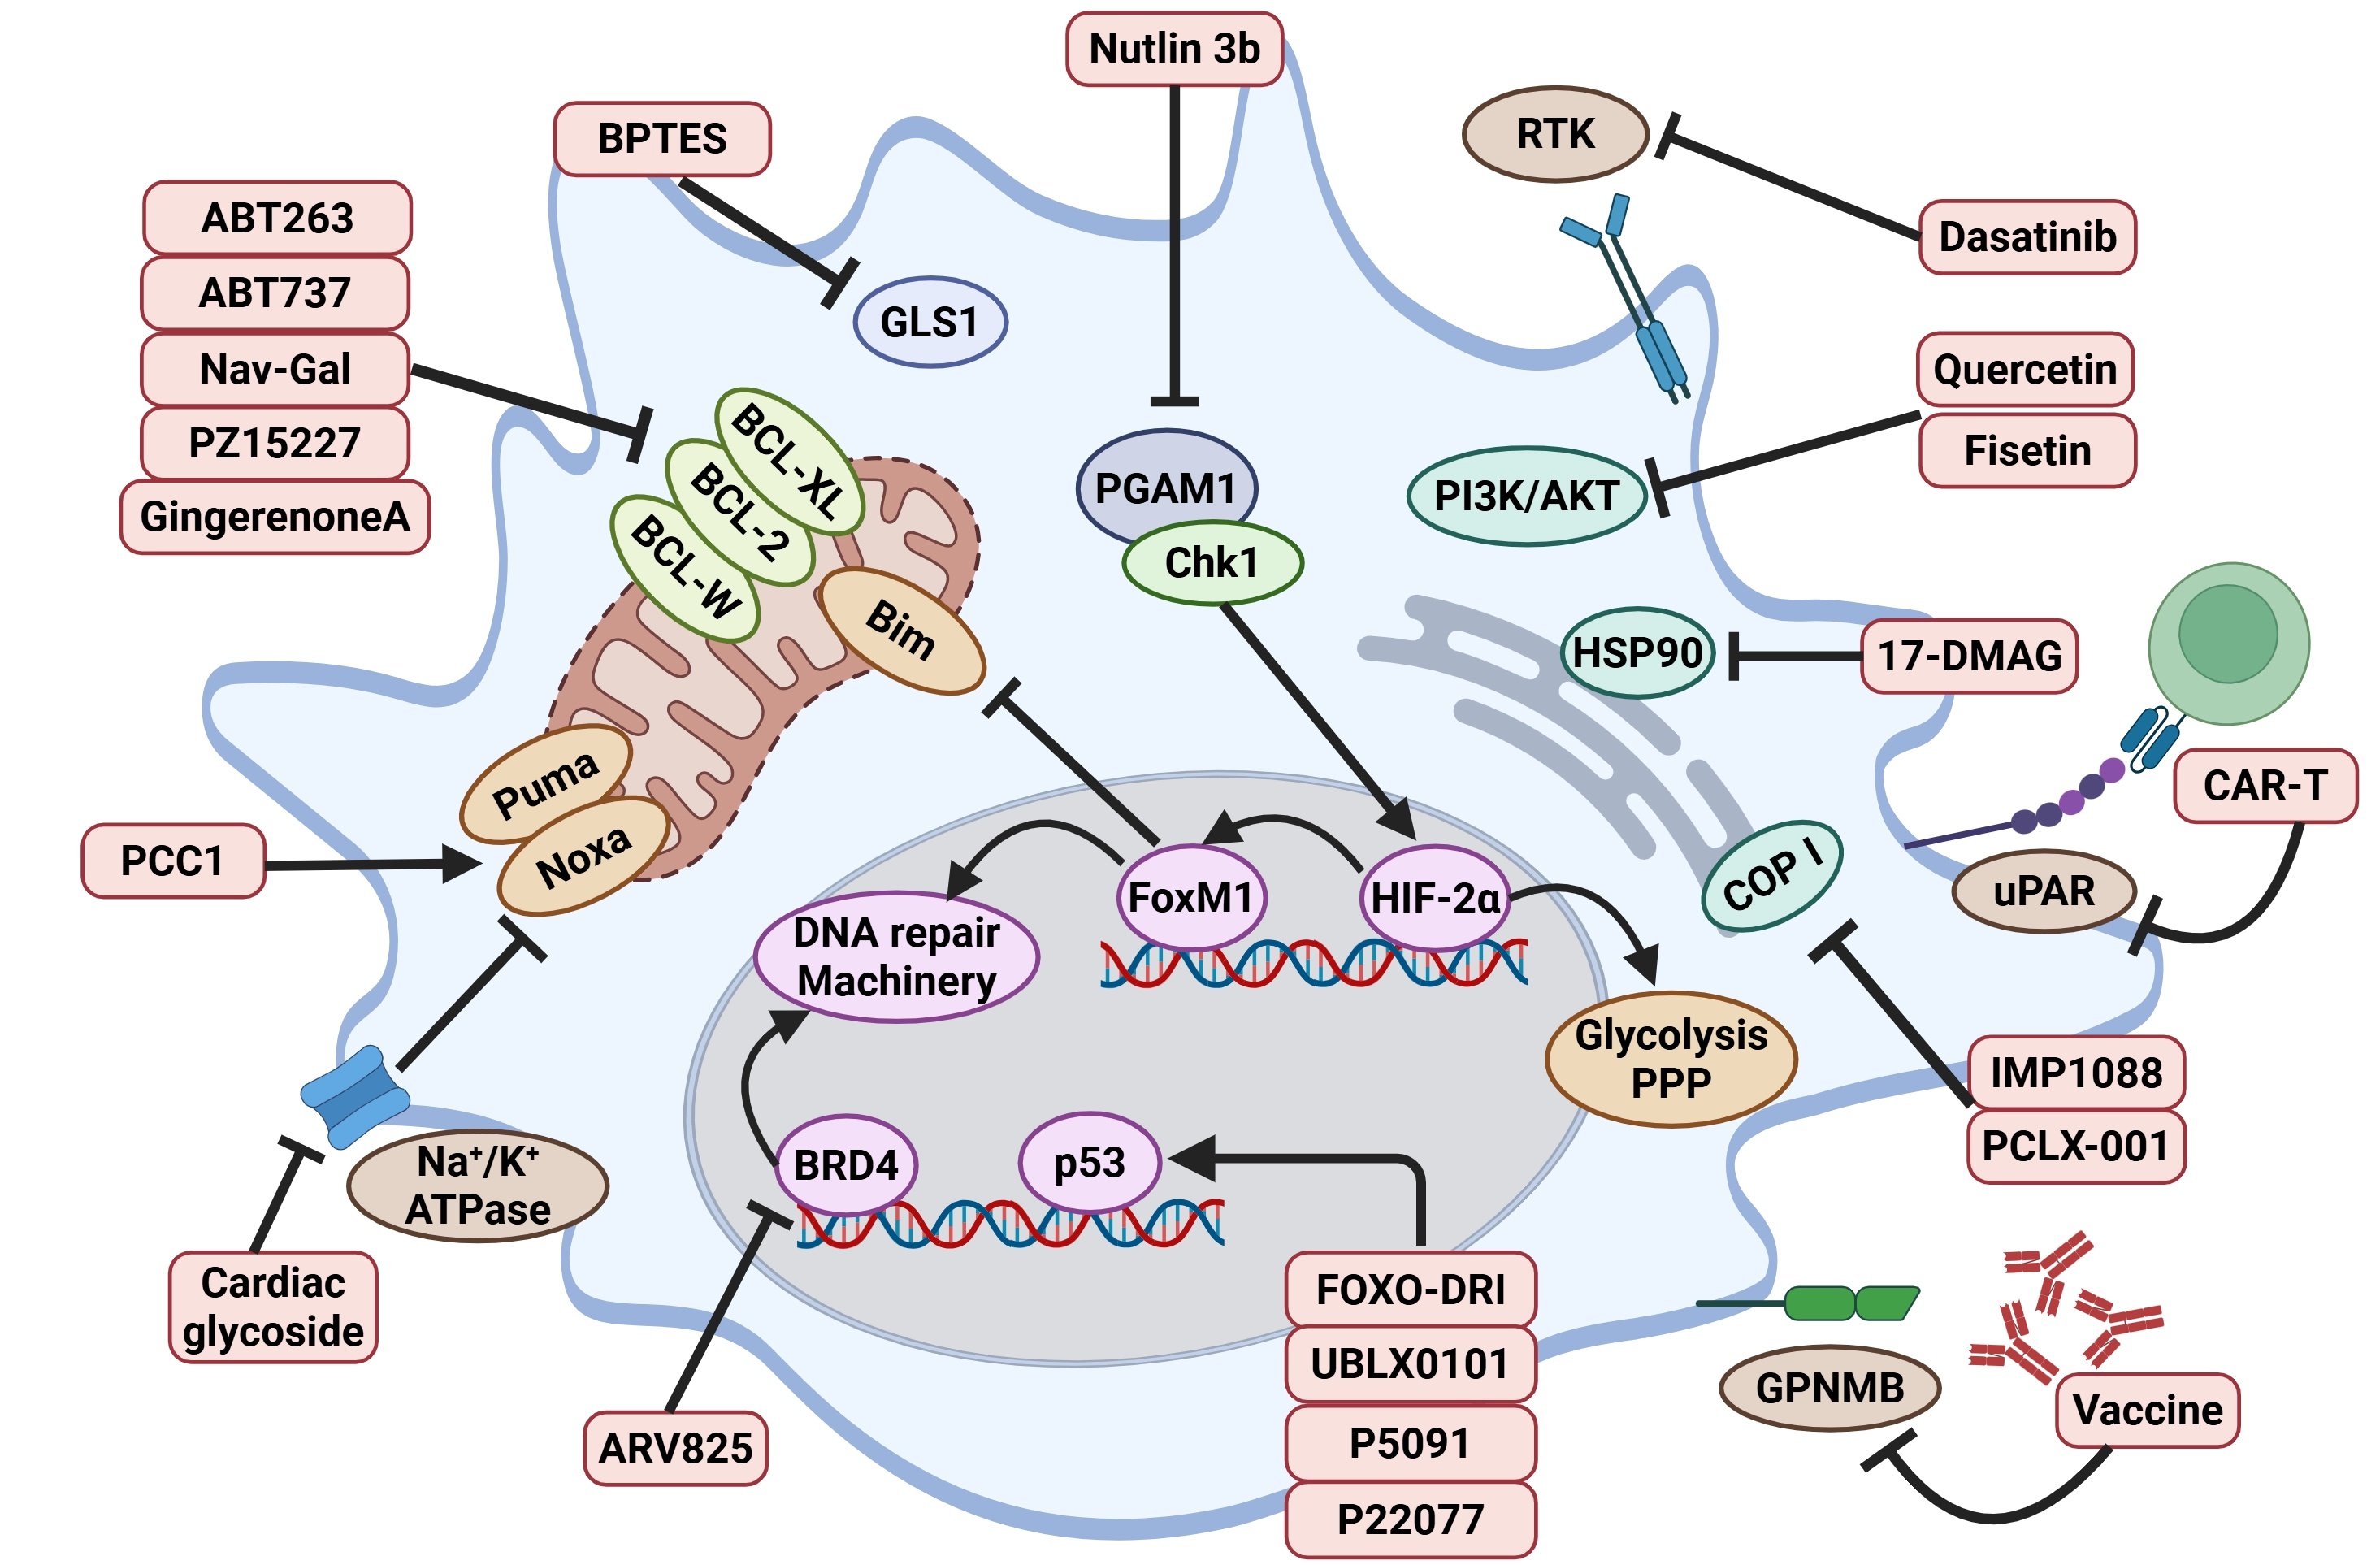
Figure 3. Overview of subclasses for senolytic approaches. Target proteins for senolysis drugs are classified into several groups. Anti-apoptotic BCL family proteins (BCL-XL, BCL-2, and BCL-W) are targeted by ABT analogues (ABT263 and ABT737), Nav-Gal, PZ15227, and Gingerenone A. Alternatively, activation of pro-apoptotic factors (Puma and Noxa) or p53 also serves as senotherapy. It is noteworthy that Nutlin 3b inhibits another proapoptotic BIM via suppression of FoxM1. Inhibitors of proteostasis proteins (HSP90 or COPI), 17-DMAG, IMP1088, and PCLX-001, are also effective for senolysis. In addition, senescence-specific membranous proteins (uPAR or GPNMB), upstream signaling modules for survival (RTK or PI3K/AKT), and metabolic enzyme (GLS1) are targeted by individual senolytic drugs or modalities. Interestingly, FoxM1 and epigenetic factor BRD4, a target of senolytic ARV825, regulate DNA repair machinery in SnCs. Created in BioRender. Kondoh, H. (2026) https://app.biorender.com/illustrations/6a4899f3caedd0741b201799. BCL: B-cell lymphoma 2; p53: tumor protein p53; FoxM1: Forkhead box M1; HSP90: heat shock protein 90; COPI: coat protein complex I; uPAR: urokinase plasminogen activator receptor; GPNMB: glycoprotein nonmetastatic melanoma protein B; RTK: receptor tyrosine kinase; PI3K/AKT: phosphoinositide 3-kinase/protein kinase; GLS1: glutaminase 1; BRD4: bromodomain-containing protein 4; SnCs: senescent cells; HIF: hypoxia-inducible factor; CAR-T: chimeric antigen receptor T cell.
4. Senolytic Selectivity of Nutlin 3b
There was no doubt that the downstream of PGAM1-Chk1 binding is FoxM1, but the challenge has resurfaced. R. Ribeiro et al. reported that intermittent activation of FoxM1 prolongs the lifespan of aged mice[49], which is seemingly inconsistent with our findings of inactivating FoxM1 by senolytic Nutlin 3b[37]. In the former study, FoxM1 was periodically activated from an early stage in young mice (8 weeks old). The mice bore the transgene of the constitutive active FoxM1 with N-terminus truncation (FoxM1-dNdK), under a tet-on system. The transgene is periodically activated through a weekly cycle of 3-days on and 4-days off by the tet “on-off” system for 80 weeks until natural aging of mice. In contrast, in our study we treated very aged mice (20 months old) with senolytic Nutlin 3b for 3 months. Senolytic Nutlin 3b effectively suppressed FoxM1 in vitro and in vivo. The contrast between the results of senolysis (inhibiting FoxM1) and Ribeiro et al. (periodically activating FoxM1) suggests the “Goldilocks” principle of FoxM1: early/cyclic activation promotes repair, but chronic accumulation at late stage supports the survival of harmful SnCs. Thus, the therapeutic benefit of FOXM1 inhibition or activation is likely to depend on the context of diseases and tissues involved. Thus, two distinct approaches, which seem to act in the opposing direction against FoxM1 at different life stages, are similarly effective as senotherapy for beneficial outcome in aged mice.
Notably, FoxM1 mRNA continuously decreases in all organs tested until middle age of mice (40 weeks old), while in old age (90 weeks old), two distinct profiles for FoxM1 mRNA are observed; (1) a re-activated pattern in aged lungs, liver, kidneys, etc., and (2) a depressed pattern in the heart, white adipose tissue (WAT), brain, etc.[37]. We observed that senolytic Nutlin 3b is effective in the former tissues, aged lungs, liver, and kidneys, but not in the latter ones, e.g. aged WAT. In line with such tissue-dependent activation of FoxM1 in aged mice, it has been reported that FoxM1 also accumulates in lung fibrosis, one of the age-related diseases. Nutlin 3b was found to have a therapeutic effect against fibrotic lungs of mice. Thus, senolytic Nutlin 3b could be more effective in FoxM1-accumulating organs, than others.
Due to the complexity and heterogeneity of senescence in vivo, a recent study suggests that preserved endothelial functions are crucial to restore the health of mice after senolytic treatment[50]. More recently, a harmful effect of senolysis, e.g. ABT-263, was reported, which reduced the plaque stability of atherosclerosis[51]. To disclose the translational potential of Nutlin 3b, we examined its senolytic effect in different subtypes of SnCs in vitro. We observed that Nutlin 3b is effective as a senolytic for senescent fibroblasts and macrophages, but not for senescent endothelial vessel cells in vitro. Similarly, D + Q treatment also preserves vessel integrity in vivo[50], although underlying mechanisms of cell-type selectivity by Nutlin 3b or D + Q remain to be uncovered. These findings support the efficacy and minimized side effects of Nutlin 3b as a senolysis drug for the treatment of aging and age-related diseases. Although Nutlin 3b has demonstrated promising senolytic activity and an acceptable safety profile in mouse models[37], its clinical feasibility in humans remains to be established. Future studies should address the current limitations of this therapeutic strategy, including pharmacokinetic properties, long-term safety, optimal dosing, tissue distribution, and the identification of biomarkers for patient stratification or treatment monitoring.
5. Conclusion
Identification of PGAM1-Chk1 binding as a property of SnCs disclosed the biological significance of the pseudo-Warburg effect in cellular senescence. This interaction stabilizes HIF-2α by preventing its degradation, to sustain the transcription of glycolytic enzymes, simultaneously enhancing PPP activity, and upregulates FoxM1, supporting the anti-apoptotic capacity of SnCs. FoxM1 promotes the expression of DNA repair machineries while repressing the proapoptotic factor Bim. Thus, the enhanced glycolysis in SnCs is not merely an energy-generating process but is tightly coupled with DNA repair, nucleotide metabolism, and anti-apoptotic signaling, forming the core survival basis of SnCs. More importantly, the soluble factor promoting PGAM1-Chk1 interaction is not classical SASP inflammatory factors, including cytokines (interleukin-6 (IL-6) and IL-1β), chemokines (C-C motif chemokine ligand 2 (CCL2) and C-X-C motif chemokine ligand 12 (CXCL12)), growth factors (amphiregulin (AREG) and osteoprotegerin (OPG), and glycolytic modulators (insulin-like growth factor-binding proteins 1-6 (IGFBP1-6)), but the glycolytic by-product lactate, suggesting that the external inflammatory and the intrinsic survival phenotype of SnCs, although related, are mechanistically distinct, with regard to the metabolic state.
Strikingly, Nutlin 3b, an isomer of Nutlin 3a, efficiently inhibits PGAM1-Chk1 binding, but not that of p53-Mdm2, serving as a senolysis drug in vitro and in vivo. Consistent with the negative effect of Nutlin 3b against FoxM1, lung fibrosis in mice is also alleviated by senolytic Nutlin 3b, as FoxM1 is a critical driver of lung fibrosis[52]. Further verification of the senolytic effect of Nutlin 3b would help its future clinical application.
Acknowledgements
We thank all staff members of the Geriatric Unit and the Department of Diabetes, Endocrinology and Nutrition at Kyoto University for their assistance. The authors are responsible for the accuracy and scientific content of the article.
Authors contribution
Mikawa T: Conceptualization, writing-original draft.
Kameda M, Zhang Z: Writing-original draft.
Kondoh H: Supervision, writing-review & editing.
Conflicts of interest
The authors declare no conflicts of interest.
Ethical approval
Not applicable.
Consent to participate
Not applicable.
Consent for publication
Not applicable.
Availability of data and materials
Not applicable.
Funding
This work was supported in part by grants from the MEXT/JSPS (23K27507 and 23H03884).
Copyright
© The Author(s) 2026.
References
-
2. Cooks T, Puzis R, Cohen O. Tissue resilience: Lessons from social resilience: Social sciences research on community resilience could inspire biology to understand homeostasis. EMBO Rep. 2021;22(8):EMBR202152926.[DOI]
-
3. Gorgoulis V, Adams PD, Alimonti A, Bennett DC, Bischof O, Bishop C, et al. Cellular senescence: Defining a path forward. Cell. 2019;179(4):813-827.[DOI]
-
4. Biran A, Zada L, Abou Karam P, Vadai E, Roitman L, Ovadya Y, et al. Quantitative identification of SnCs in aging and disease. Aging Cell. 2017;16(4):661-671.[DOI]
-
5. López-Otín C, Blasco MA, Partridge L, Serrano M, Kroemer G. Hallmarks of aging: An expanding universe. Cell. 2023;186(2):243-278.[DOI]
-
8. Valieva Y, Ivanova E, Fayzullin A, Kurkov A, Igrunkova A. Senescence-associated β-galactosidase detection in pathology. Diagnostics. 2022;12(10):2309.[DOI]
-
10. Serrano M, Lin AW, McCurrach ME, Beach D, Lowe SW. Oncogenic ras Provokes Premature Cell Senescence Associated with Accumulation of p53 and p16INK4a. Cell. 1997;88(5):593-602.[DOI]
-
11. Huang W, Hickson LJ, Eirin A, Kirkland JL, Lerman LO. Cellular senescence: The good, the bad and the unknown. Nat Rev Nephrol. 2022;18(10):611-627.[DOI]
-
12. Wu Z, Qu J, Liu GH. Roles of chromatin and genome instability in cellular senescence and their relevance to ageing and related diseases. Nat Rev Mol Cell Biol. 2024;25(12):979-1000.[DOI]
-
16. Passos JF, Saretzki G, Ahmed S, Nelson G, Richter T, Peters H, et al. Mitochondrial dysfunction accounts for the stochastic heterogeneity in telomere-dependent senescence. PLoS Biol. 2007;5(5):e110.[DOI]
-
19. Coppé JP, Patil CK, Rodier F, Sun Y, Muñoz DP, Goldstein J, et al. Senescence-associated secretory phenotypes reveal cell-nonautonomous functions of oncogenic RAS and the p53 tumor suppressor. PLoS Biol. 2008;6(12):e301.[DOI]
-
22. Birch J, Gil J. Senescence and the SASP: Many therapeutic avenues. Genes Dev. 2020;34(23-24):1565-1576.[DOI]
-
24. Chaib S, Tchkonia T, Kirkland JL. Cellular senescence and senolytics: The path to the clinic. Nat Med. 2022;28(8):1556-1568.[DOI]
-
27. Zhao H, Liu Z, Chen H, Han M, Zhang M, Liu K, et al. Identifying specific functional roles for senescence across cell types. Cell. 2024;187(25):7314-7334.e21.[DOI]
-
30. Zhao J, Tan C, Imai R, Ukon N, Shimoyama S, Maejima Y, et al. Evaluation of organ glucose metabolism by 18F-FDG accumulation with insulin loading in aged mice compared with young normal mice. Sci Rep. 2021;11:7421.[DOI]
-
31. Liu B, Meng Q, Gao X, Sun H, Xu Z, Wang Y, et al. Lipid and glucose metabolism in senescence. Front Nutr. 2023;10:1157352.[DOI]
-
32. Warburg O. On respiratory impairment in cancer cells. Science. 1956;124(3215):269-270.[PubMed]
-
33. Li Z, Munim MB, Sharygin DA, Bevis BJ, Vander Heiden MG. Understanding the Warburg effect in cancer. Cold Spring Harb Perspect Med. 2025;15(12):a041532.[DOI]
-
34. Mikawa T, Shibata E, Shimada M, Ito K, Ito T, Kanda H, et al. Phosphoglycerate mutase cooperates with Chk1 kinase to regulate glycolysis. iScience. 2020;23(7):101306.[DOI]
-
35. Dixon AS, Schwinn MK, Hall MP, Zimmerman K, Otto P, Lubben TH, et al. NanoLuc complementation reporter optimized for accurate measurement of protein interactions in cells. ACS Chem Biol. 2016;11(2):400-408.[DOI]
-
38. TeSlaa T, Ralser M, Fan J, Rabinowitz JD. The pentose phosphate pathway in health and disease. Nat Metab. 2023;5(8):1275-1289.[DOI]
-
45. McHugh D, Durán I, Gil J. Senescence as a therapeutic target in cancer and age-related diseases. Nat Rev Drug Discov. 2025;24(1):57-71.[DOI]
-
46. Rad AN, Grillari J. Current senolytics: Mode of action, efficacy and limitations, and their future. Mech Ageing Dev. 2024;217:111888.[DOI]
-
49. Ribeiro R, Macedo JC, Costa M, Ustiyan V, Shindyapina AV, Tyshkovskiy A, et al. In vivo cyclic induction of the FOXM1 transcription factor delays natural and progeroid aging phenotypes and extends healthspan. Nat Aging. 2022;2(5):397-411.[DOI]
-
52. Penke LR, Speth JM, Dommeti VL, White ES, Bergin IL, Peters-Golden M. FOXM1 is a critical driver of lung fibroblast activation and fibrogenesis. J Clin Invest. 2018;128(6):2389-2405.[DOI]
Copyright
© The Author(s) 2026. This is an Open Access article licensed under a Creative Commons Attribution 4.0 International License (https://creativecommons.org/licenses/by/4.0/), which permits unrestricted use, sharing, adaptation, distribution and reproduction in any medium or format, for any purpose, even commercially, as long as you give appropriate credit to the original author(s) and the source, provide a link to the Creative Commons license, and indicate if changes were made.
Publisher’s Note
Science Exploration remains a neutral stance on jurisdictional claims in published
maps
and institutional affiliations. The views expressed in this article are solely those
of
the author(s) and do not reflect the opinions of the Editors or the publisher.
Share And Cite


Science Exploration Style
Mikawa T, Kameda M, Zhang Z, Kondoh H. PGAM1-Chk1 interaction as a novel target for senolysis to preserve resilience during aging. Geromedicine. 2026;2:202625. https://doi.org/10.70401/Geromedicine.2026.0034
Tips
Copy completed.
Submit a Manuscript
Author Instructions
Cite this Article
Article Metrics
0
View
0
Download
Cited
Article Updates

- Abstract
- Keywords
- 1. Introduction
- 2. Biological Significance of Increased Glycolytic Metabolism in SnCs
- 3. Senolytic Mechanism by Inhibition of PGAM1-Chk1 Binding
- 4. Senolytic Selectivity of Nutlin 3b
- 5. Conclusion
- Acknowledgements
- Authors contribution
- Conflicts of interest
- Ethical approval
- Consent to participate
- Consent for publication
- Availability of data and materials
- Funding
- References
- Copyright
 Get citation
Science Exploration Style

Mikawa T, Kameda M, Zhang Z, Kondoh H. PGAM1-Chk1 interaction as a novel target for senolysis to preserve resilience during aging. Geromedicine. 2026;2:202625. https://doi.org/10.70401/Geromedicine.2026.0034
copy
Share Link
copy