Bridging the scales: Advanced scanning probe microscopy across molecular redox regimes

Noah Al-Shamery
1,*

,
Rafael A. Vicente
2
,
Christian Zocher
3
,
Berthold Kersting
3
,
Kirill Monakhov
1,*
*Correspondence to:
Noah Al-Shamery, Leibniz Institute of Surface Engineering (IOM), Leipzig 04318, Germany.
E-mail: noahcoelestin.alshamery@leibniz-iom.de
Kirill Monakhov, Leibniz Institute of Surface Engineering (IOM), Leipzig 04318, Germany. E-mail: kirill.monakhov@iom-leipzig.de
Kirill Monakhov, Leibniz Institute of Surface Engineering (IOM), Leipzig 04318, Germany. E-mail: kirill.monakhov@iom-leipzig.de
Smart Mater Devices. 2027;3:202646. 10.70401/smd.2026.0054
Received: July 30, 2026Accepted: October 09, 2026Published: October 12, 2026
This article belongs to the Special lssue Intelligent Soft and Organic Matter: Fundamentals, Electrochemistry, and Sensing
Abstract
Redox-active ligands, metal complexes, and larger coordination frameworks are chemically precise, responsive building blocks for smart materials, molecular electronics, energy storage, and neuromorphic device concepts. However, their redox behavior depends strongly on the experimental regime in which it is probed. Bulk electrochemistry, confined nanodroplet methods, electrochemical scanning probe microscopy, tunneling spectroscopy, and synchrotron-coupled nanospectroscopy often probe the same molecular event through fundamentally different physical observables. In this mini-review, we use polyoxometalates as a guiding molecular class to compare redox behavior across three length-scale regimes: bulk ensembles, confined nanoscale assemblies, and single-molecule or few-molecule junctions. We discuss cyclic voltammetry, scanning electrochemical probe methods, electrochemical scanning tunneling microscopy, scanning tunneling spectroscopy, electron-spin-resonance scanning tunneling microscopy (ESR-STM), scattering-type near-field optical microscopy, and synchrotron nanospectroscopic methods. We argue that advanced scanning probe microscopy provides the missing conceptual and experimental bridge between classical electrochemical redox potentials and nanoscale electronic charge and spin structure, enabling a regime-aware design language for redox-programmable molecular materials.
Keywords
Scanning probe microscopy, electrochemistry, polyoxoanions, SECCM, EC-STM, s-SNOM, smart materials, solution-processable electronics
1. Introduction
Redox-active organic and inorganic molecules are central to the design of smart materials and devices because they combine discrete charge and spin states, chemical tunability, and external addressability by voltage, light, ions, solvent, pH, or local fields. Polyoxometalates (POMs) are particularly appealing and useful representatives of this class of responsive compound types: they are atomically precise molecular metal-oxide clusters, typically anionic and charge-balanced by countercations, that can undergo reversible multi-electron redox transformations over broad potential windows without necessarily losing structural integrity[1]. This makes them worthwhile for electrocatalysis, charge storage, molecular electronics, resistive switching, and neuromorphic
Rather than representing a complication, this regime dependence highlights the need to combine complementary information obtained across different experimental length scales, with advanced scanning probe microscopy serving as an important bridge between them. Classical electrochemistry provides averaged thermodynamic and kinetic information over large molecular populations. Scanning electrochemical probe methods introduce liquid confinement and spatial addressability. EC-STM and scanning tunneling spectroscopy (STS) provide sub-molecular structural and electronic readout at interfaces[7]. Electron-spin-resonance STM (ESR-STM) extends this logic toward spin-resolved molecular states[8]. Optical and synchrotron-coupled near-field methods, including scattering-type scanning near-field optical microscopy (s-SNOM), synchrotron infrared nanospectroscopy, nanoscale Fourier transform infrared spectroscopy (nano-FTIR), and related beamline-based surface probes, add vibrational, chemical, and electronic contrast beyond the faradaic current[9]. The aim of this mini-review is therefore not to introduce a single new apparatus, but to develop a regime-aware framework for redox-active molecular systems. We keep POMs as the conceptual “glue” because they are chemically robust, strongly redox-active, ionic, and relevant across solution, surface, and solid-state environments. At the same time, the discussion naturally extends to other redox-active molecular materials, including coordination complexes, molecular electrolytes, ionic liquids, redox polymers, small conducting polymers like eumelanin, molecular perovskites, and hybrid
We organize the review across three length-scale regimes: bulk ensembles, confined nanoscale ensembles, and single-molecule or few-molecule junctions. Across these regimes, we ask how advanced scanning probe microscopy (SPM) and beamline-coupled nanoscale spectroscopy can connect classical electrochemical observables to local structure, oxidation state, conductance, spin, and chemical identity. As a focused Mini-Review rather than a systematic review, the literature was selected to represent experimentally established examples spanning bulk electrochemistry, confined electrochemical probe methods, tunneling-based measurements, and nanospectroscopy, with particular emphasis on POMs and closely related redox-active molecular systems. Recent literature was prioritized together with foundational methodological studies and representative primary reports. The novelty of the present synthesis lies in organizing these otherwise separate literatures according to experimental regime and explicitly comparing how molecular population, environment, electrode coupling, and observable change across scales.
2. Why Redox-Active Molecular Systems Require a Regime-Aware Framework
The simplest electrochemical system consists of a redox molecule as a species that exchanges electrons with an electrode at a defined potential. In bulk solution, this picture is powerful because diffusion, electrolyte composition, and electrode area can often be described using continuum models. For POMs, CV can reveal multiple reduction and oxidation waves, apparent reversibility, proton coupling, stability windows, and kinetic limitations. These quantities remain foundational for any device-oriented discussion[12,13]. Yet molecular materials intended for smart devices rarely operate as ideal dissolved species. They may be adsorbed, trapped in thin films, patterned in microdroplets, incorporated into polymer matrices, stabilized by counterions, deposited onto conducting surfaces, or integrated into nanoscale junctions. In such cases, the molecular redox state becomes coupled to confinement, hydration, interfacial fields, and local electronic structure. A redox event may appear as a faradaic peak in CV, a local current transient in SECCM, an apparent height or contrast change in EC-STM, a shift in local density of states in STS, a vibrational fingerprint in nano-IR, or a conductance/spin transition in tunneling-based readout[14].
At first glance, these measurements may appear to describe different physical phenomena. In favorable cases, these measurements can report on different consequences of the same underlying redox process, but such correspondence must be established experimentally rather than assumed from similar potential- or bias-dependent signatures. Assignment of a tunneling or conductance feature to a defined redox transition is strongest when its potential dependence can be correlated with an independently established electrochemical redox window and when alternative contributions from electrode coupling, molecular configuration, counterion redistribution, and electrostatic gating can be excluded or constrained.
This creates a conceptual discontinuity between electrochemistry and molecular electronics. Electrochemists often think in terms of redox potentials, electrolyte, mass transport, and faradaic charge. Molecular electronics often thinks in terms of energy-level alignment, tunneling barriers, metal-molecule coupling, and current-voltage characteristics. Advanced SPM provides one route between these languages by combining local structural/electronic interrogation with chemically controlled environments, depending on the specific technique employed[15-17]. For POMs, this bridge is especially important because their multi-electron redox chemistry and charge-compensating counterions are sensitive to local environment. Hydration may stabilize reduced states; ion pairing may shift apparent potentials; adsorption may distort geometry or alter electronic coupling; and drying may freeze a
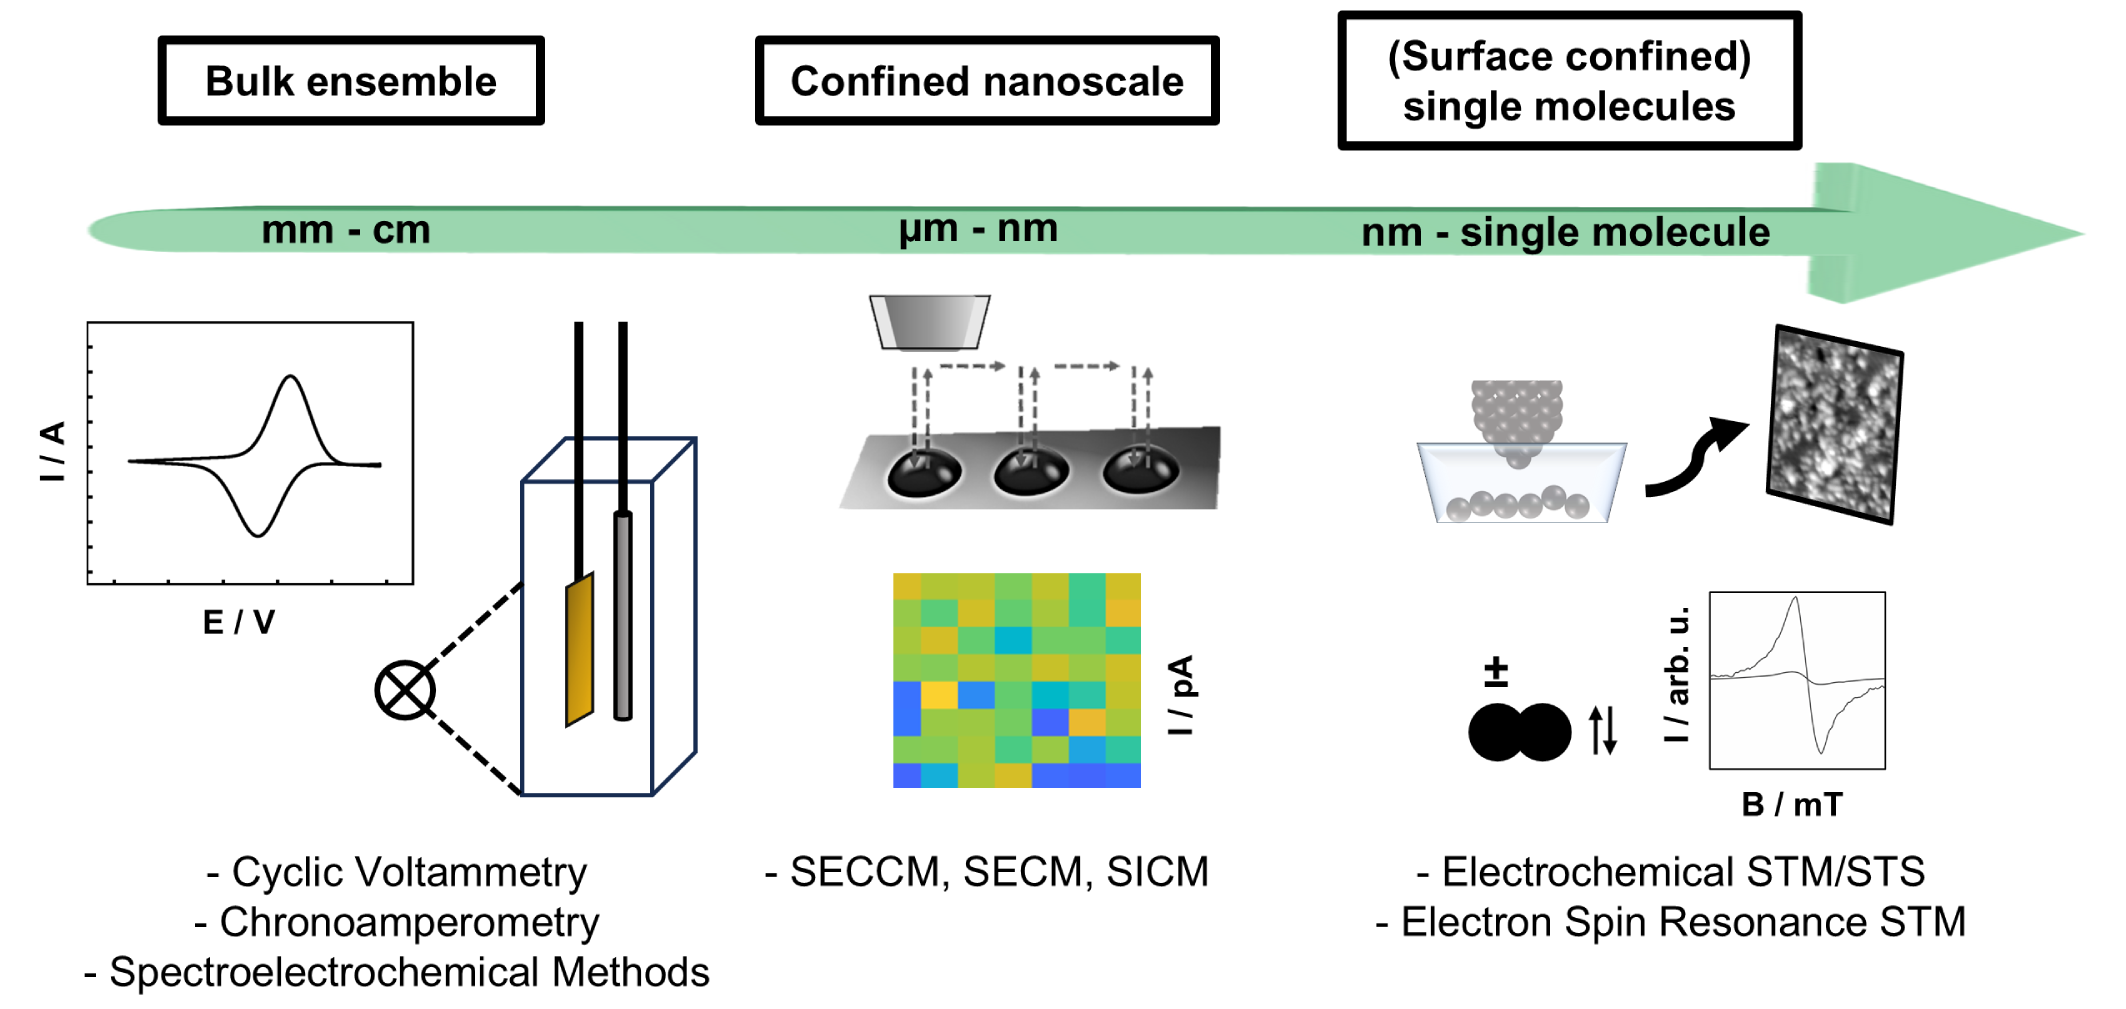
Figure 1. Regime map for redox-active molecular systems across length scales. Conceptual comparison of bulk ensemble electrochemistry, confined nanodroplet electrochemistry, and single-molecule or few-molecule tunneling regimes. Bulk CV and spectroelectrochemistry define the reference redox window and ensemble stability. SECCM, SICM, and SECM introduce liquid-on-solid confinement and spatially localized faradaic readout. EC-STM, STS, STM, and ESR-STM probe surface-confined molecules through tunneling, local density of states, and potentially spin-sensitive signals. Moving from left to right, the regimes reflect a progressive transition from
3. Bulk Ensemble Regime
Bulk electrochemistry remains the reference regime for redox-active molecular systems. Techniques such as CV, differential pulse voltammetry, chronoamperometry, and spectroelectrochemistry establish the baseline redox window, reversibility, charge-transfer kinetics, and chemical stability of a molecular compound. For POMs, these measurements are particularly informative because multiple metal centers can participate in stepwise electron accumulation, often accompanied by proton or counterion effects[19]. In the bulk ensemble regime, the strengths are reproducibility, quantitative current response, and straightforward comparison across molecular libraries. This is the regime in which one can rapidly screen POM structures, countercations, solvents, ionic strength, and pH. It also allows comparisons to other redox-active species, such as viologens, quinones, ferrocenes, molecular catalysts, coordination complexes, and ionic-liquid-based electrolytes[23,24].
The limitation is that bulk electrochemistry averages over many molecules and many local environments. Surface adsorption, electrode heterogeneity, aggregation, local precipitation, and redox-induced restructuring may be hidden within an apparently
Bulk spectroelectrochemistry partially addresses this problem by correlating electrochemical state with optical, ultraviolet
Thus, bulk electrochemistry should be viewed as the first layer of a regime-aware workflow. It defines the chemical redox map. Advanced SPM then asks which parts of that map survive, shift, broaden, fragment, or become electronically useful when the same molecules are confined at surfaces or integrated into nanoscale junctions.
4. Confined Nanoscale Ensembles
Between bulk solution and single-molecule tunneling lies an experimentally intermediate regime: the confined liquid-on-solid nanodroplet or microdroplet. Scanning electrochemical probe microscopy (SEPM), including SECCM, scanning ion conductance microscopy (SICM), and scanning electrochemical microscopy (SECM), provides spatially localized electrochemical access to surfaces while retaining a liquid environment. In SECCM, a nanopipette confines electrolyte and redox-active species in a meniscus that contacts a substrate, creating a local electrochemical cell whose size can range from micrometers down to the nanoscale[5,28]. For POMs, this regime is compelling because it combines confined liquid-phase electrochemistry with localized electrochemical interrogation of surfaces. A droplet can deliver a defined molecular population to a selected surface location, apply a programmed electrochemical waveform, and then retract, leaving behind a transient or dried molecular domain. In this sense, SECCM is not only an analytical technique but also a local redox-programming and deposition tool[3,29,30].
This capability is significant for smart materials and devices. Instead of globally coating a substrate with a redox-active film, confined droplet methods can create arrays of chemically distinct or redox-distinct surface pixels. Each pixel may experience a different potential history, molecular concentration, solvent composition, or dwell time. For molecular ionic compounds such as POMs, such redox-programmed domains could encode charge state, spin state, catalytic activity, or chemical memory[31,32]. However, confined droplet electrochemistry is not simply miniaturized bulk electrochemistry. The droplet geometry, evaporation rate, meniscus lifetime, ion migration, concentration gradients, wetting, surface roughness, and local conductance all influence the measured response. A POM that appears reversible in bulk solution may undergo adsorption, local enrichment, ion-pairing, or precipitation in a nanodroplet[33,34].
SICM and SECM provide complementary perspectives. SICM is powerful for topographical and ion-conductance feedback, especially at soft, biological, or insulating interfaces. SECM is widely used for redox mediator feedback, catalytic mapping, and local reactivity. SECCM is particularly attractive for creating local electrochemical cells on surfaces and for high-throughput mapping. For the present mini-review, these methods are best treated as a family of liquid-confined electrochemical SPM tools rather than as competing communities[35].
The key message is that confined nanodroplets create an ensemble regime that is neither bulk nor single molecule. The molecule count is reduced, the substrate matters strongly, and the local redox state can be deliberately controlled through the applied electrochemical potential. This regime is where POMs and other ionic molecular compounds can be translated from solution chemistry into spatially addressable surface function. For device-oriented applications, successful local redox writing must also be distinguished from non-volatile state retention. A state generated within a biased liquid meniscus may relax after removal of the potential, evaporation of the electrolyte, counterion redistribution, or structural reorganization during drying. Consequently, retention time, reversibility over repeated write/read cycles, and preservation of molecular identity across the wet-to-dry transition are critical benchmarks when translating SECCM-written redox states toward memory or switching architectures.
5. Single-Molecule and Few-Molecule Regimes: EC-STM, STM/STS, and ESR-STM
At the smallest length scales, redox-active molecules are interrogated as individual entities or few-molecule ensembles. STM and
STS extends this approach by measuring local current-voltage characteristics and differential conductance, thereby probing local electronic states and energy-level alignment. For redox-active molecular systems, tunneling-based spectroscopy and transport measurements can reveal how electrochemical changes in molecular charge state influence electronic structure, energy-level alignment, and charge transport[40]. Here, solvent and electrolyte are removed or drastically reduced, and the molecule is probed through direct electronic coupling to electrodes. For POMs and related ionic compounds, this raises fundamental questions: Which redox states remain stable after drying? Are counterions and residual water part of the functional junction? Does an applied bias reproduce a redox transition, or does it induce a non-equilibrium electronic response? These questions are central to molecular memory and switching concepts and should be explored in detail in the future to obtain a complete understanding of such condensed matter[41-43]. It is important to distinguish these measurements from electrochemically gated break-junction experiments. In EC-STM and STS, tunneling primarily probes a surface-confined adsorbate and its local electronic structure, whereas a break junction measures charge transport through a molecule coupled between two electrodes. Moreover, electrochemical potential controls the interfacial redox environment and molecular charge state, while source-drain bias drives non-equilibrium transport across the junction. Consequently, a potential-dependent conductance change is not by itself proof of a molecular redox event: junction geometry, electrode-molecule coupling, energy-level alignment, and electrostatic gating must also be considered. Correlation with an independently established redox window therefore provides a substantially stronger assignment than conductance changes alone. For this context, the term ‘single-molecule regime’ therefore describes the number of molecular entities interrogated rather than a single experimental geometry or common electrical observable.
ESR-STM adds a further dimension: spin sensitivity at the level of individual atoms or molecules. ESR-STM extends conventional STM by combining atomic-scale tunneling measurements with electron spin resonance, enabling the coherent detection and manipulation of individual electron spins on surfaces. In most current implementations, a radio frequency (RF) excitation is applied to the tunneling junction while a spin-polarized STM tip detects changes in the tunneling current associated with resonant spin transitions. Because spin coherence is readily disrupted by thermal fluctuations and environmental interactions, these experiments are typically performed under ultra-high vacuum at cryogenic temperatures (≈ 1-4 K) and in the presence of an external magnetic field, often using thin insulating decoupling layers to preserve the magnetic properties of the adsorbed species. Although technically demanding, this combination of atomic spatial resolution with electron-spin sensitivity provides a unique opportunity to directly relate molecular electronic structure, oxidation state, and magnetic properties within a single nanoscale experiment. ESR-STM is conceptually important for redox-active single molecules and assemblies because many redox events generate or quench unpaired spins. Although ESR-STM has not yet been established for POM redox systems, demonstrations on atomic and molecular spin systems suggest a possible future route toward correlating local charge, spin, and electronic structure in appropriately designed redox-active POMs[44,45].
More broadly, electrochemical control provides an additional degree of freedom for engineering molecular spin systems by modifying oxidation state, electronic configuration, and local electrostatic environment. Such capabilities may ultimately enable studies of redox-dependent spin transport, spin polarization, and hyperpolarization mechanisms in molecular systems. This
6. Advanced Optical and Synchrotron-Coupled SPM
While conventional spectroscopic methods are widely used in conjunction with bulk electrochemistry to probe ensemble-averaged behavior, synchrotron-based techniques open the door to powerful nanospectroscopic approaches for investigating materials at the nanoscale[46]. The high brilliance and tunability of synchrotron radiation enable the use of techniques such as synchrotron infrared nanospectroscopy (SINS) for nanoscale vibrational spectroscopy and chemical identification beyond the diffraction limit[47]. X-ray microscopy approaches, including X-ray nanodiffraction, spatially-resolved X-ray absorption spectroscopy, and X-ray fluorescence microscopy, can provide spatially resolved information on local structure, oxidation state, ligand environment, and elemental composition[48,49]. Collectively, these synchrotron-based methods provide a versatile platform for visualizing and characterizing nanoscale structural, chemical, and electronic phenomena that are difficult to resolve using conventional spectroscopic or microscopy techniques, enable the visualization of features hidden by ensemble averaging, and provide a more comprehensive understanding of spatially heterogeneous processes at the nanoscale[50].
In contrast, advanced optical methods can also provide nanoscale information through the interaction of light with molecular, electronic, and vibrational degrees of freedom, often with greater accessibility and experimental flexibility. Although conventional optical microscopy is diffraction-limited, approaches such as conventional laser-based s-SNOM, tip-enhanced Raman spectroscopy (TERS), and photothermal microscopy can achieve nanoscale or sub-diffraction-limited spatial information while simultaneously providing chemical and functional contrast[51].
It is possible to infer the chemical identity of redox-active molecules using fingerprints extracted at confined ensembles and
These methods could serve as powerful validation tools for redox-active molecular systems and may be adapted to POMs and related ionic molecular materials. This capability is particularly valuable for validating POM thin films, SECCM-written microdroplet arrays, and redox-programmed surfaces, where nanoscale heterogeneity and local chemical changes can critically influence functionality. More broadly, synchrotron nanospectroscopy can be applied to ionic molecular materials and molecular electronic devices to correlate molecular-scale redox processes with device-relevant structure and performance, thereby building an important bridge between fabrication, characterization, and functional validation.
7. Toward Regime-Bridging Molecular Analysis
A summary of selected techniques discussed in this mini-review that could be and have been applied to POMs and other related compounds is depicted in Figure 2. The field now needs a shared comparison framework for comparing redox-active molecular systems across regimes. Here, “regime-bridging” does not imply direct numerical equivalence between observables obtained by different techniques. Rather, it denotes a comparison framework in which the molecular identity and redox state are referenced as consistently as possible while the experimental environment, molecular population, electrochemical control, and measured observable are explicitly recorded. Cross-regime correspondence is strongest when the same redox transition can be independently assigned in more than one regime; it becomes uncertain when confinement, drying, electrode coupling, counterion redistribution, or non-equilibrium bias changes the molecular state itself. A useful workflow would begin with bulk electrochemistry to establish the molecular redox map, proceed to confined nanodroplet methods to test local programmability, use EC-STM and STS to connect redox state with surface electronic structure, and then apply optical or synchrotron-coupled nanospectroscopy to verify chemical
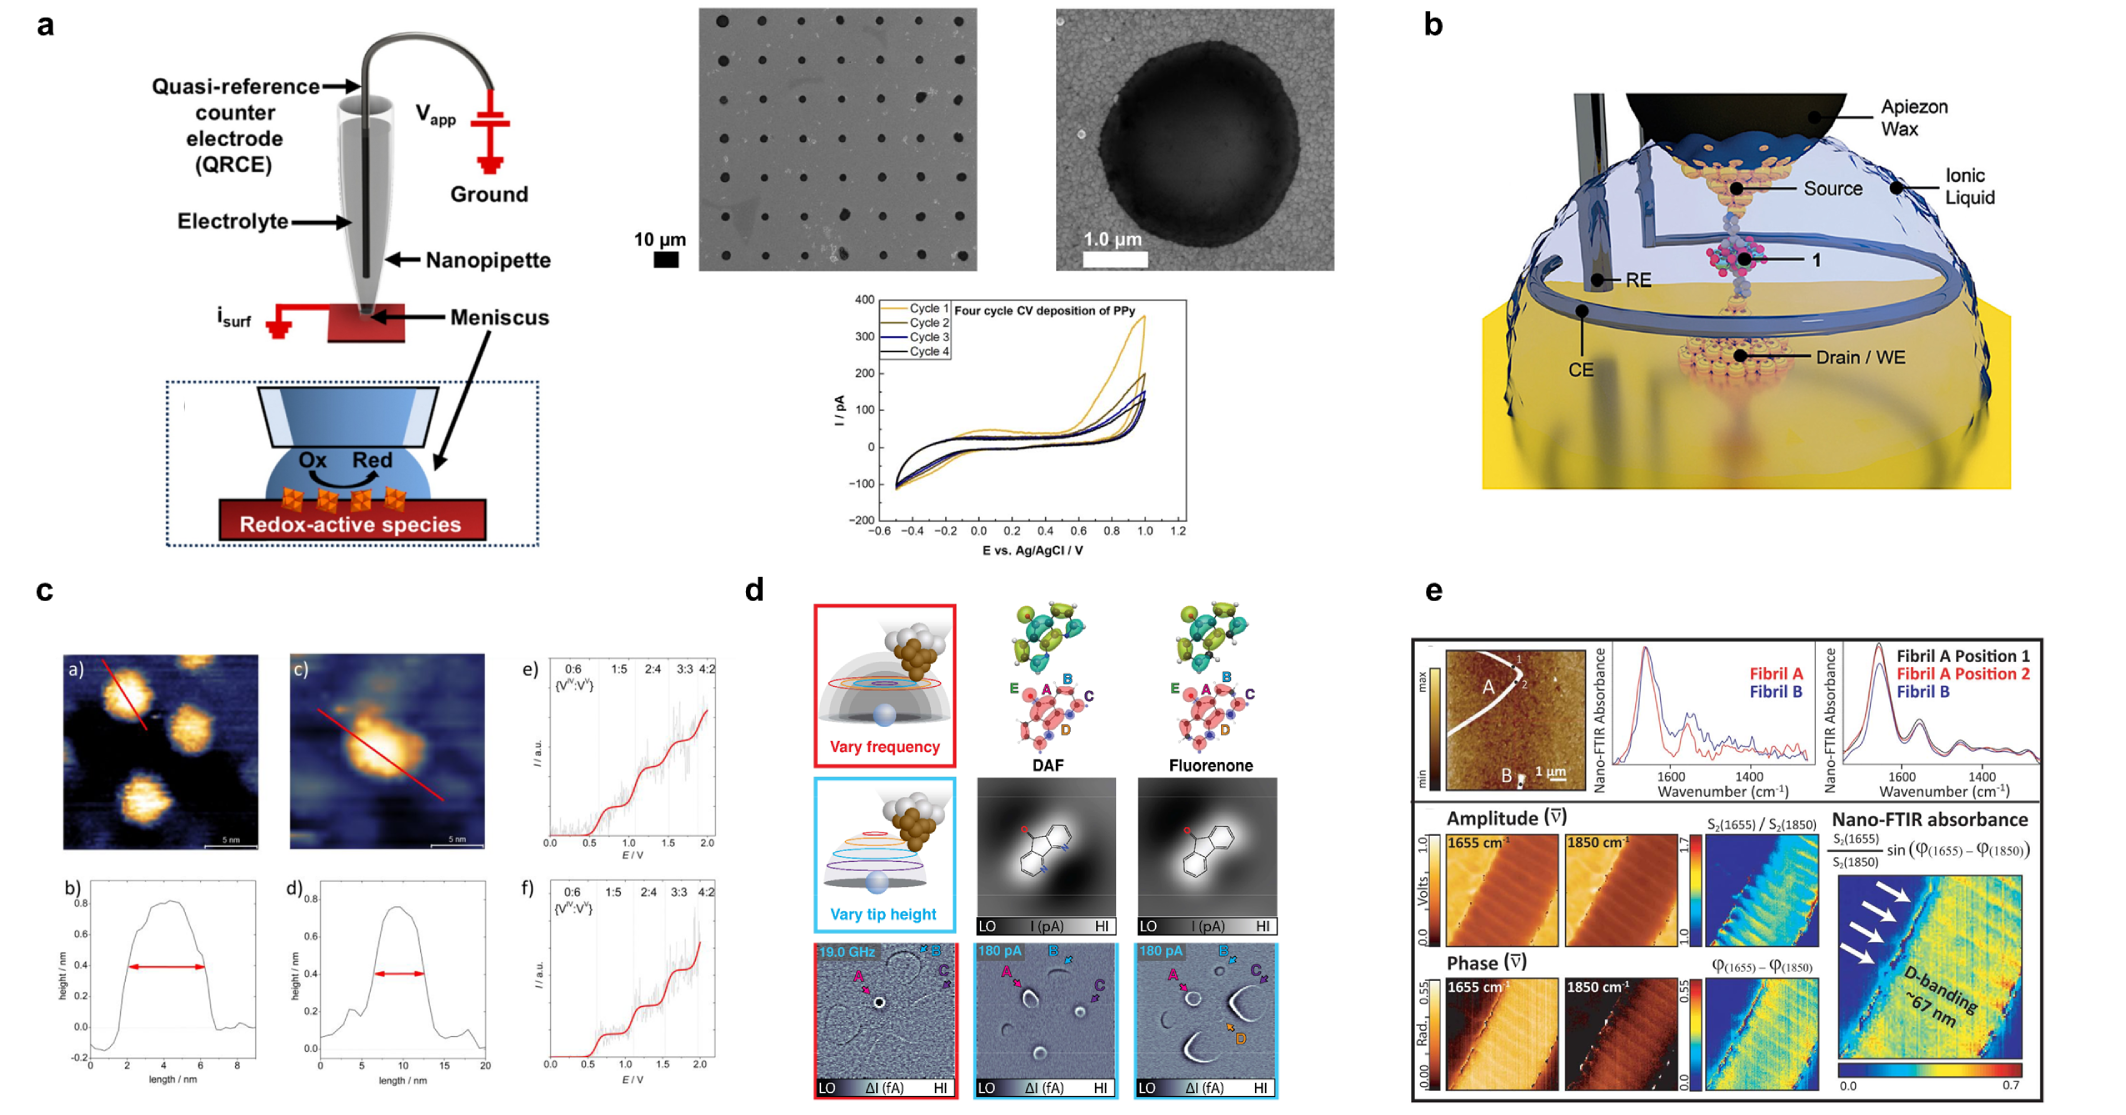
Figure 2. Representative advanced SPM and nanospectroscopy toolset that could be and have been used for POM-based redox materials and other related compounds; where direct POM examples are unavailable, non-POM methodological examples are explicitly indicated. Selected examples of techniques relevant to regime-bridging redox analysis: (a) SECCM for confined electrochemical patterning: Using nanopipettes for the redox programming of POMs and for the 2D-patterning of polypyrrole (PPy) nanodeposits using cyclic voltammetry. Reproduced from references[3,4]. CC BY 4.0; (b) EC-STM for molecular imaging at interfaces: the shown example is a single-POM junction [MnMo6O18{(OCH2)3CC5H4N}2]3- in a four-electrode set-up immersed in an ionic liquid. Reproduced from references[17]; Reproduced with permission from
The value of such cross-regime analysis ultimately depends on the intended application. For sensing, local measurements can reveal heterogeneity relevant to sensitivity, selectivity, drift, and response time; for memory or switching concepts, retention, endurance, switching variability, and energetic cost become central; and for energy-storage materials, charge retention and cycling stability are more relevant. These benefits must be balanced against practical limitations of nanoscale methods, including throughput, instrument access, sample preparation, and possible probe-induced perturbation. Regime-bridging measurements should therefore be selected according to the device-relevant question rather than applied as a fixed analytical sequence.
8. From POMs to Related Ionic and Redox-Active Molecular Materials
Although POMs are central to this case study, the regime-aware framework is broadly applicable to other redox-active molecular systems. Coordination complexes, for example, can exhibit well-defined metal- and ligand-centered redox states and are widely studied for application in molecular electronics, catalysis, spin crossover, and light-driven switching[58,59]. Although these material classes differ substantially in chemical composition, dimensionality, and electronic structure, they share a dependence on reversible charge transfer and its coupling to the local chemical environment. The same regime-aware framework can therefore be used to examine how redox behavior evolves from bulk electrochemistry to confined interfaces and ultimately to single-molecule measurements. Comparing these regimes can reveal how ligand design influences electrode coupling, electronic structure, and redox-state stability[58,60].
A key starting point in such systems is the choice of the redox-active metal center. Among the transition-metal ions commonly investigated in electrochemical and magneto-spectroscopic studies, copper and iron provide particularly instructive examples. Copper can switch between the closed-shell d10 Cu(I) state (S = 0) and the open-shell d9 Cu(II) state (S = 1/2), providing an accessible one-electron spin-charge transition[61]. Iron spans a broader range of electronic and spin configurations. Fe(II), for example, can adopt low-spin (S = 0) or high-spin (S = 2) configurations depending on the ligand field, while Fe(III) can occur in low-spin (S = 1/2) and high-spin (S = 5/2) states[62]. The electronic and magnetic properties of these ions are well established, and their redox chemistry can be tuned within potential windows compatible with a broad range of organic ligands. However, as isolated ions or simple aqua complexes, they do not provide the structural and electronic control required for switchable, addressable, and spatially localized spin-charge functionality. The metal ion provides the redox-active center, but the surrounding coordination environment determines how that redox event can be coupled to a functional device. To transform a metal ion into a functional and addressable molecular unit, a defined ligand framework is therefore required[63-68].
Among bio-inspired coordination scaffolds, porphyrins provide an archetypal example. Nature has extensively exploited this structural motif in chlorophyll, hemoglobin, melanin-like tetramer systems, and cytochromes[69-71]. Porphyrinoid motifs involved in biological electron-transfer pathways further illustrate the versatility of this design principle[72,73]. Incorporation of a metal ion into the porphyrin framework imposes a defined coordination geometry and provides a conjugated electronic structure that couples the metal d orbitals to the frontier orbitals of the macrocycle. The resulting complex therefore links redox state, spin multiplicity, and electronic coupling to the surrounding environment and, ultimately, to an electrode[74].
In their closed-shell oxidation or reduction states, many such complexes are diamagnetic and can be addressed electrochemically through controlled potential coulometry at the respective redox potential. As the applied potential traverses the corresponding redox window, the system can enter an open-shell, paramagnetic state. At the single-molecule level, such a transition can become a discrete charge-transfer event, in which the addition or removal of a single electron changes the molecular spin state and can simultaneously modify tunneling conductance, the local density of states, and spin-resonance signals[75]. In a confined nanodroplet, the same transition is sampled over a small ensemble of molecules, and the measured current reflects the corresponding ensemble population. In bulk solution, the cyclic voltammogram integrates the response of a much larger molecular population, while spin information is accessed through complementary ensemble techniques such as EPR or SQUID measurements. The same underlying redox-spin process can therefore appear as a discrete molecular transition, a localized current response, or a macroscopic voltammetric wave, depending on the experimental regime. This illustrates a central principle of the regime-aware framework: the molecular redox chemistry may remain the same, while the observable and its sensitivity to the local environment change with the measurement scale.
Scaling from single molecules and small ensembles to macroscopic materials, individual spin-redox units can also be incorporated into extended three-dimensional architectures. Conducting-polymer composites provide one example in which nanoscale organization and connectivity can translate local electronic functionality into macroscopic conduction through percolating
Across these systems, the same question arises as those posed throughout this review: How does an ensemble redox map established by one technique translate to crystalline or surface-confined environments, and which redox and spin states remain accessible? Can a verified single-molecule spin transition be translated into a measurable macroscopic magnetic or conductive response? Addressing these questions requires the cross-regime comparison enabled by advanced SPM and beamline-coupled nanospectroscopy. Such comparisons can establish which molecular properties are intrinsic to the redox-active unit, which depend on confinement and electrode coupling, and which emerge only at larger length scales. The regime-aware framework therefore generalizes at the level of experimental variables rather than identical molecular behavior. Confinement, electrode coupling, local electrostatics, solvation, and measurement timescale are relevant across many redox-active molecular systems, whereas strong counterion coupling, highly accessible multi-electron reduction, cluster rigidity, or specific spin-state manifolds are material-dependent. Cross-regime comparisons should therefore preserve these distinctions rather than assuming that behavior established for POMs transfers quantitatively to coordination complexes, polymers, or other ionic materials. Representative examples illustrating this progression from molecular redox and spin states to surface-confined, single-molecule, and extended functional materials are summarized in Figure 3.
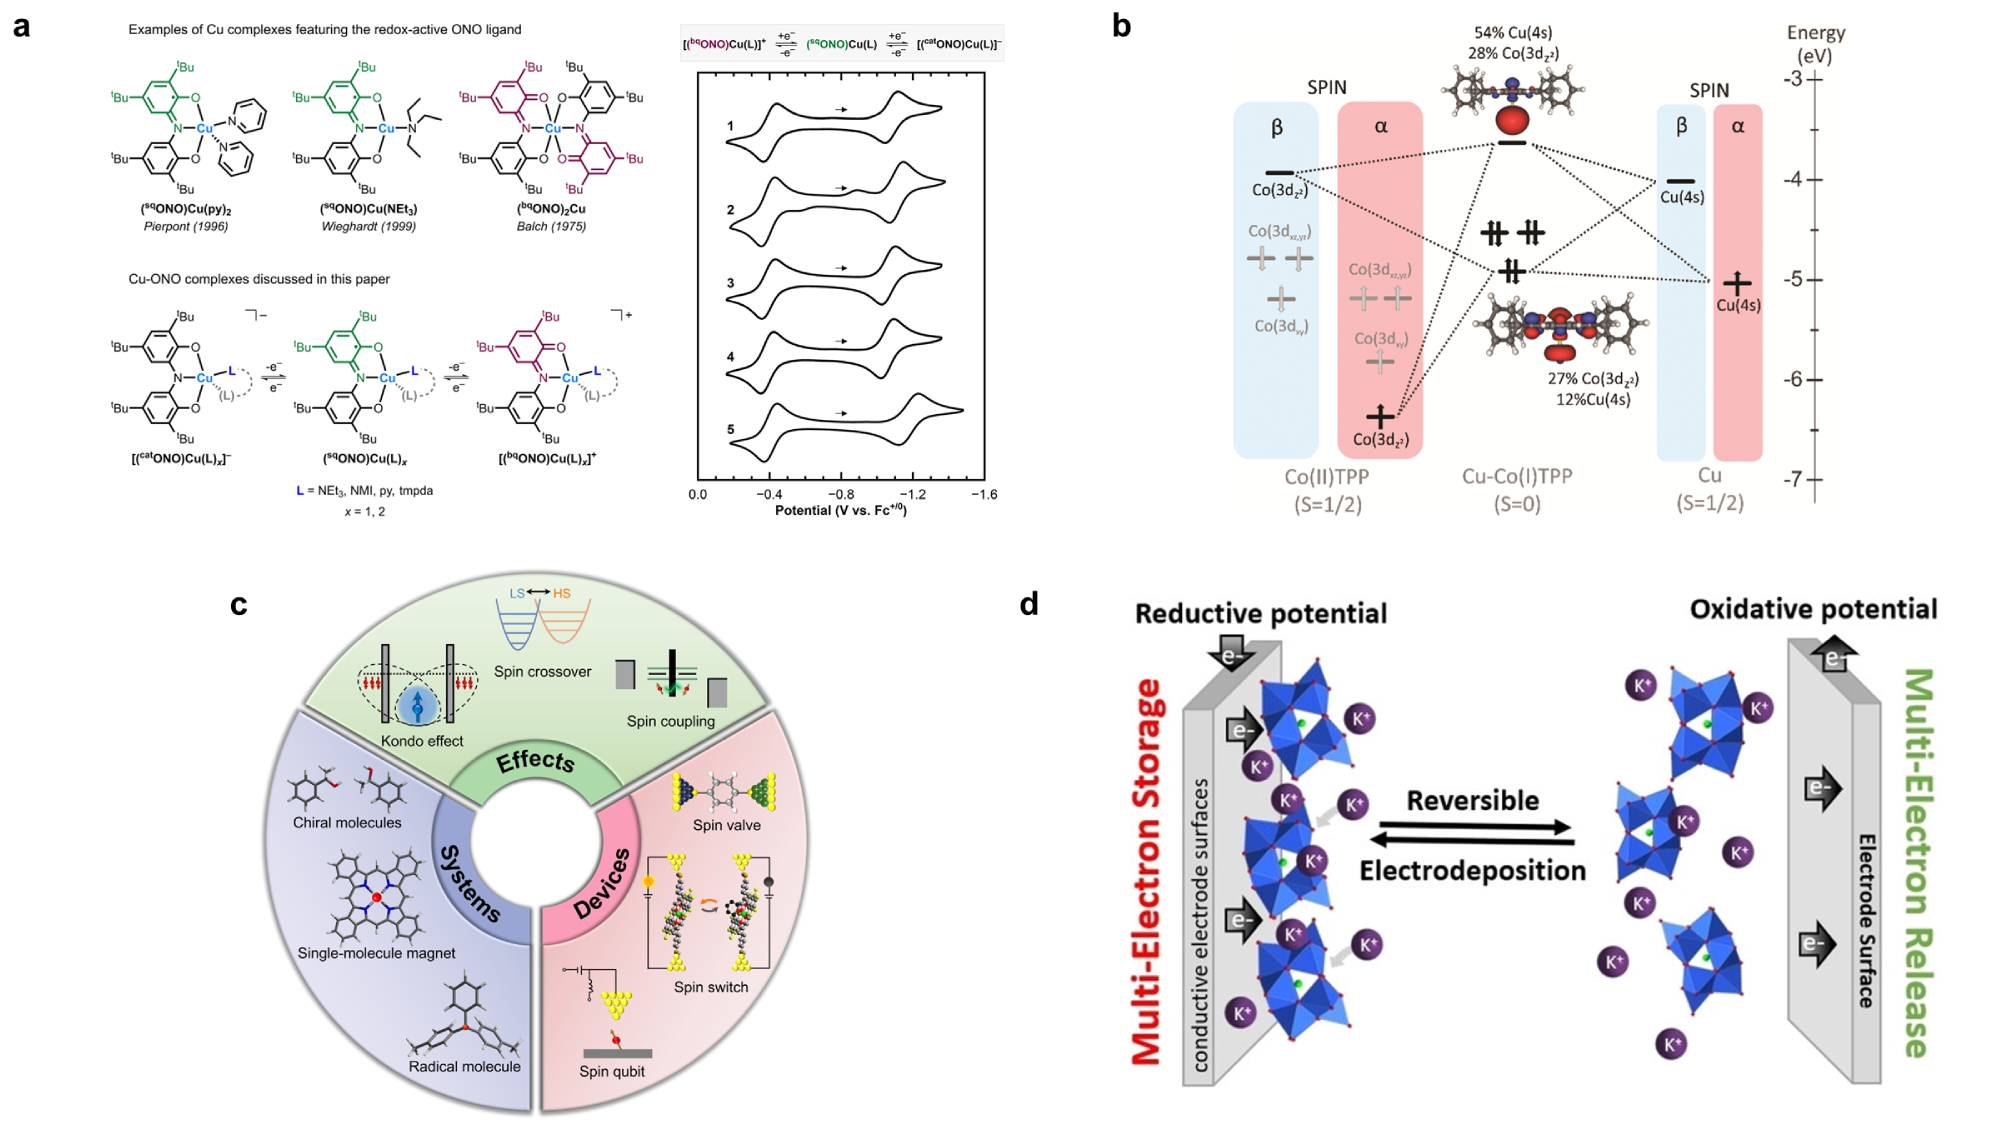
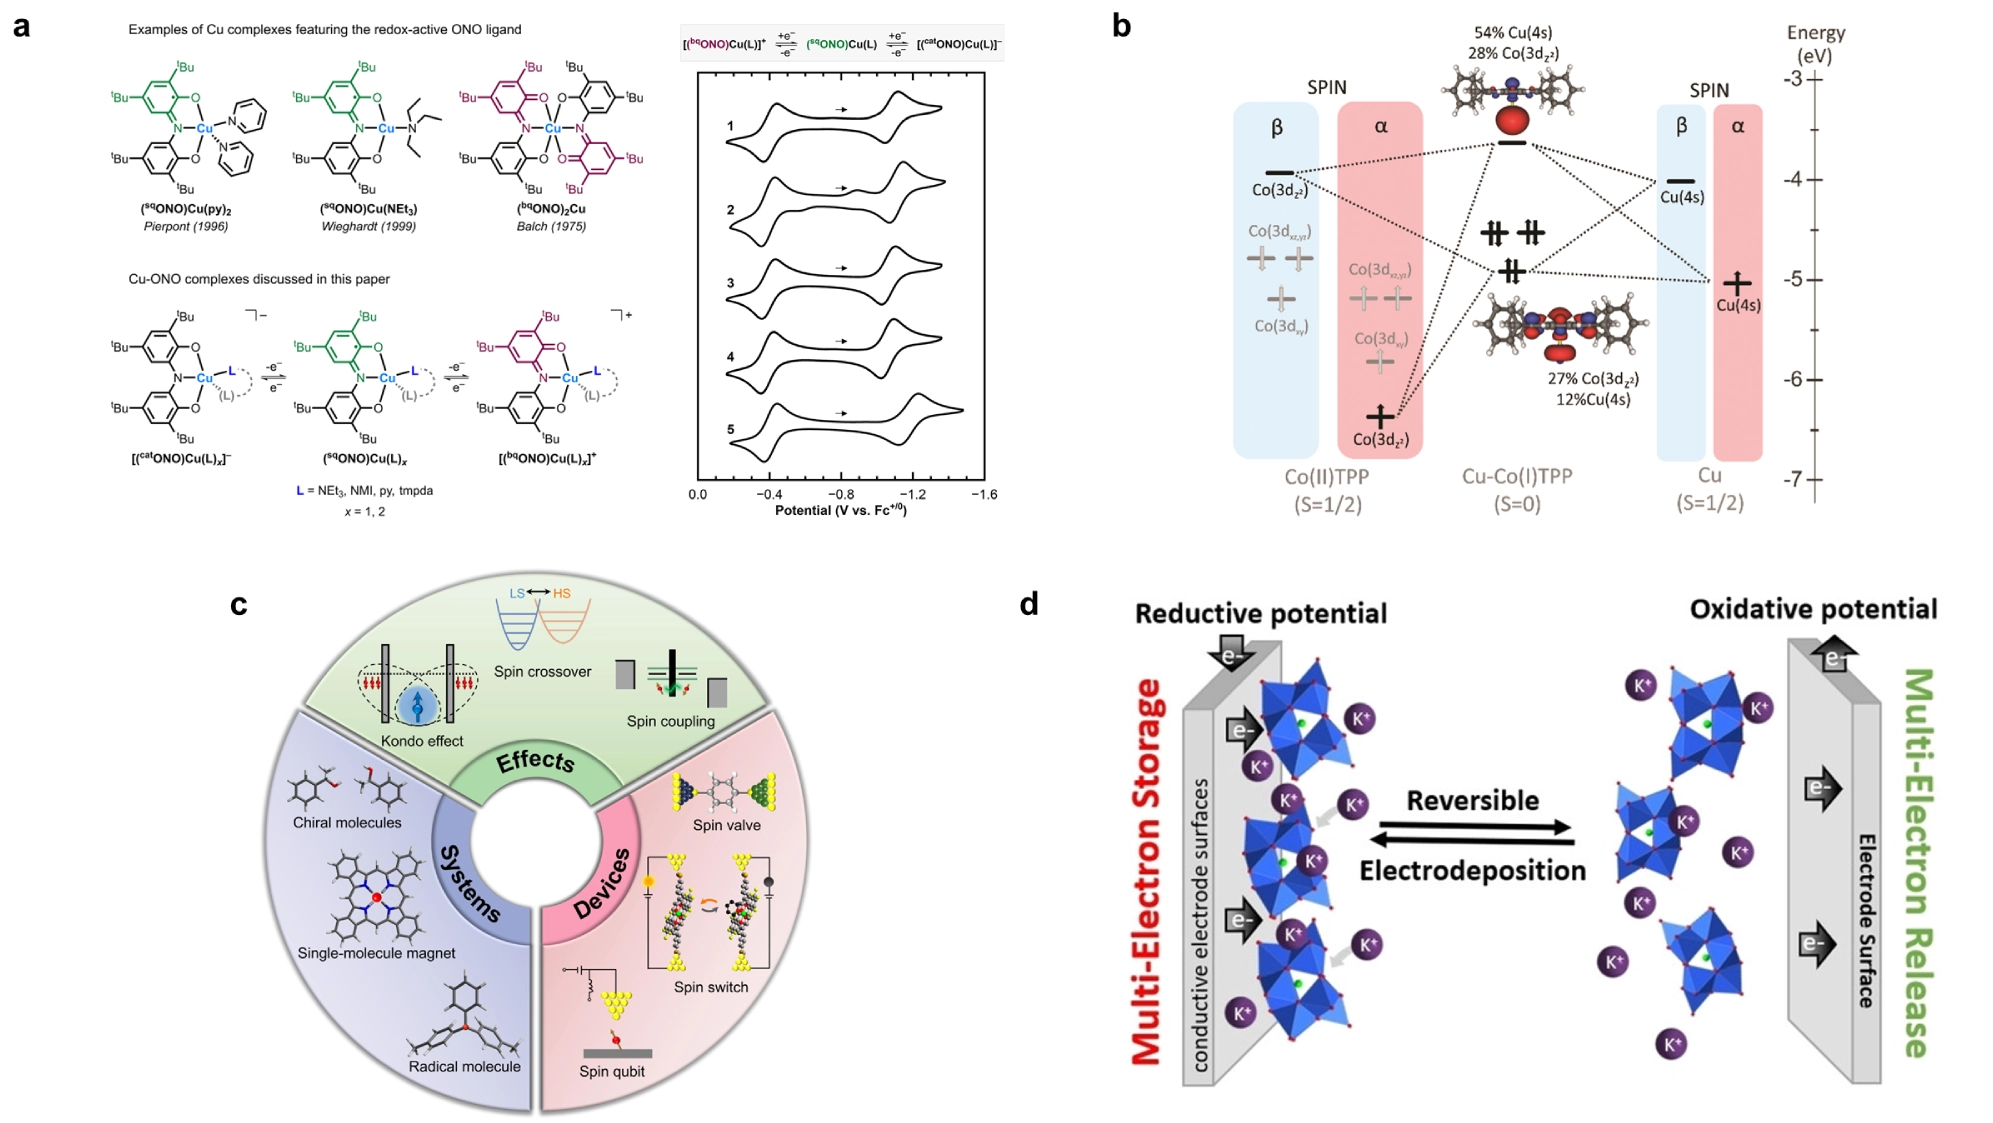
Figure 3. Representative redox-active molecular systems illustrating the extension of the regime-aware framework beyond individual POM examples. (a) Coordination complexes with redox-active metal-ligand units provide chemically defined molecular oxidation states and tunable electronic structures. Reproduced from reference[68]. CC BY 4.0; (b) Surface-supported metalloporphyrins illustrate the coupling between molecular oxidation state, spin state, and electrode environment. Reproduced from reference[74]. CC BY-NC 3.0; (c) Single-molecule spin devices extend this concept toward molecular-scale electronic and spin readout, where discrete charge and spin states can be interrogated at the individual-molecule level. Reproduced from reference[75]. CC BY 4.0; (d) Molecular vanadium oxides illustrate the translation of discrete,
9. Conclusion
Redox-active molecular systems display different behavior depending on whether they are measured as bulk ensembles, confined nanodroplet ensembles, or individual surface-bound molecules (summary in Table 1). POMs exemplify this regime dependence because their multi-electron redox chemistry is strongly influenced by hydration, counterions, spatial confinement at interfaces, and electrode coupling. Bulk electrochemistry defines the thermodynamic and kinetic reference state; SECCM, SICM, and SECM introduce liquid-confined spatial addressability; EC-STM and STS connect redox state to tunneling and local electronic structure; ESR-STM points toward spin-resolved redox comparison frameworks; and s-SNOM, synchrotron nano-IR, X-ray photoelectron spectroscopy (XPS), and X-ray absorption spectroscopy (XAS) provide chemical validation beyond current readout. Together, these complementary approaches illustrate how advanced SPM enables a regime-bridging view of molecular electrochemistry, with POMs serving as a representative model system. Beyond molecular electronics, this regime-aware framework may also provide new opportunities for exploring redox-controlled spin phenomena, where electrochemical control over molecular charge states can be coupled to
Table 1. Regime comparison for redox-active molecular systems.
| Regime/analytical role | Typical methods | Molecular population | Typical environment | Main observables | Selected strengths |
| Bulk ensemble | CV, chronoamperometry, bulk spectroelectrochemistry | Very large ensemble | Solution electrolyte | Faradaic current, optical/spectral response | Quantitative redox information |
| Confined nanoscale | SECCM, SICM, SECM | Local ensemble | (Meniscus-)confined electrolyte | Local current, local reactivity, specific surface domains | Spatially addressable redox writing/reading |
| Surface-confined molecules | EC-STM/STS | Few molecules to monolayer | Solid-liquid interface | Tunneling current, apparent height, local electronic states | Structural/electronic correlation |
| Solid-supported single/few molecules | STM/STS, ESR-STM | Single/few molecules | Vacuum, dry/controlled environment | Local density of states, spin resonance | Molecular-scale electronic/spin readout |
| Cross-regime validation | s-SNOM, nano-FTIR, XPS/XAS, synchrotron techniques | Nanoscale domains to ensembles | Dry/ambient, vacuum, or operando-compatible cells | Vibrational, chemical, oxidation states | Confirms molecular identity and state |
CV: cyclic voltammetry; SECCM: scanning electrochemical cell microscopy; SICM: scanning ion conductance microscopy; SECM: scanning electrochemical microscopy;
Acknowledgments
The authors thank colleagues involved in discussions on redox-programmed molecular surfaces and advanced scanning probe microscopy.
During the preparation and revision of this manuscript, OpenAI ChatGPT (GPT-6) was used to assist with language editing and textual refinement. All scientific content, literature selection, interpretation, and conclusions were critically evaluated, verified, and approved by the authors. No AI tools were used for data generation or data processing. The authors reviewed, revised, and approved the final manuscript and take full responsibility for its content.
Authors contribution
Al-Shamery N: Conceptualization, writing-original draft, writing-review & editing.
Vicente RA, Zocher C: Resources, writing-review & editing.
Kersting B: Resources, supervision, writing-review & editing.
Monakhov K: Conceptualization, supervision, writing-review & editing.
All authors critically reviewed and edited the manuscript, approved the final version, and agree to take responsibility for its content.
Conflicts of interest
Noah Al-Shamery serves as a Guest Editor for the Special Issue to which this manuscript was submitted and had no role in the peer-review or decision-making process for this manuscript. The other authors declare no conflicts of interest.
Ethical approval
Not applicable.
Consent to participate
Not applicable.
Consent for publication
Not applicable.
Availability of data and materials
Not applicable.
Funding
This work was supported by the Deutsche Forschungsgemeinschaft (DFG) through the Collaborative Research Center (Sonderforschungsbereich)/Transregio (SFB/TRR 386) on “Hyperpolarization in Molecular Systems – Polarization, Transport, Reactivity” (HYP*MOL, project number 514664767). N. Al-Shamery also acknowledges funding from the DFG through the Walter Benjamin Programme (project number 588571146).
Copyright
© The Author(s) 2026.
References
-
1. Monakhov K, Petrovskii SK, Long DL, Nyman M, Kahnt A, Warneke J, et al. Polyoxometalates for health, electronics, energy, and the environment: Challenges, opportunities, and future horizons. ChemistryEurope. 2026;4(5):e70280.[DOI]
-
2. Ueda T. Electrochemistry of polyoxometalates: From fundamental aspects to applications. ChemElectroChem. 2018;5(6):823-838.[DOI]
-
3. Al-Shamery N, Lee PS, Unwin PR, Monakhov K. Redox-programmed surfaces via scanning electrochemical cell microscopy (SECCM). Trends Chem. 2025;7(9):494-496.[DOI]
-
4. Al-Shamery N, Valavanis D, Dean B, Dettlaff A, Sobaszek M, Bogdanowicz R, et al. Optimizing electrochemical microprinting of conducting polymers: Scanning electrochemical cell microscopy (SECCM) coupled with conveyor-belt surface analysis. Small Methods. 2026;10:e01781.[DOI]
-
7. Filoni C, Wandelt K, Marfori L, Leone M, Duò L, Ciccacci F, et al. A combined EC-STM and EC-AFM investigation of the sulfate adsorption on a Cu(111) electrode surface up to the anodic corrosion potential. Appl Surf Sci. 2023;611:155542.[DOI]
-
9. Hillenbrand R, Abate Y, Liu M, Chen X, Basov DN. Visible-to-THz near-field nanoscopy. Nat Rev Mater. 2025;10(4):285-310.[DOI]
-
11. Kim J, Kim JH, Ariga K. Redox-active polymers for energy storage nanoarchitectonics. Joule. 2017;1(4):739-768.[DOI]
-
12. Yamasaki N, Azuma SY, Hasegawa T, Ogo S, Ueda T. Voltammetric behaviour of Keggin-type copper-substituted tungstosulfate in acetonitrile: Electron transfer processes with deposition and stripping. J Electroanal Chem. 2026;1010:120036.[DOI]
-
14. Petrangolini P, Alessandrini A, Navacchia ML, Capobianco ML, Facci P. Electron transport properties of single-molecule-bearing multiple redox levels studied by EC-STM/STS. J Phys Chem C. 2011;115(40):19971-19978.[DOI]
-
15. Keat CJ. The new frontier of nanoengineering, an atomic-level blueprints for future devices. Integr Sci Adv. 2025;1(1):13-19.[DOI]
-
16. Benstetter G, Biberger R, Liu D. A review of advanced scanning probe microscope analysis of functional films and semiconductor devices. Thin Solid Films. 2009;517(17):5100-5105.[DOI]
-
17. Wu C, Qiao X, Robertson CM, Higgins SJ, Cai C, Nichols RJ, et al. A chemically soldered polyoxometalate single-molecule transistor. Angew Chem Int Ed. 2020;59(29):12029-12034.[DOI]
-
18. Zhao Q, Li B, Menon D, Zhu C, Alizadeh Kiapi MR, Chen X, et al. Cation-directed assembly and sequential functionalization enable superprotonic polyanion–organic frameworks for high-power fuel cells. Nat Chem. 2026;18(7):1231-1241.[DOI]
-
19. Monakhov K. Stimuli-responsive gold–polyoxometalate multielectron transformers. Inorg Chem Front. 2025;12(2):396-399.[DOI]
-
20. Salhi J, Mattera M, Arfaoui I, Calupitan JP, Alves S, Troufflard C, et al. Single polyoxometalate-based nanoclusters characterized by infrared absorption nanospectroscopy. Commun Chem. 2024;7:299.[DOI]
-
21. Moors M, Monakhov KY. Multistate switchable polyoxometalates as neuromimetic emulators. Mater Today. 2024;81:1-3.[DOI]
-
23. Chang S, Chen Y, Zhang J, Pan Q, Hua Y. Heterogeneous polyoxometalate-based catalysts for functionalized quinones synthesis: Systematic advances, prospects, and challenges. ChemPlusChem. 2025;90(8):e202500261.[DOI]
-
25. Yu SH, Hassan SZ, So C, Mingyun K, Chung DS. Molecular-switch-embedded solution-processed semiconductors. Adv Mater. 2023;35(4):2203401.[DOI]
-
26. Sun L, Diaz-Fernandez YA, Gschneidtner TA, Westerlund F, Lara-Avila S, Moth-Poulsen K. Single-molecule electronics: From chemical design to functional devices. Chem Soc Rev. 2014;43(21):7378-7411.[DOI]
-
27. Zhai Y, Zhu Z, Zhou S, Zhu C, Dong S. Recent advances in spectroelectrochemistry. Nanoscale. 2018;10(7):3089-3111.[DOI]
-
29. Torres D, Bailly J, Bernal M, Coelho LB, Ustarroz J. Electrochemical nucleation and the role of the surface state: Unraveling activity distributions with a cross-system examination and a local electrochemistry approach. J Solid State Electrochem. 2024;28(5):1719-1734.[DOI]
-
30. Martín-Yerga D, Unwin PR, Valavanis D, Xu X. Correlative co-located electrochemical multi-microscopy. Curr Opin Electrochem. 2023;42:101405.[DOI]
-
31. Ghoshal A, Kaur R, Sanku S, Singh AK, Mondal PC. A twist in the molecular memory function: Chemical compositions of different redox couples control the resistive switching bias polarity. J Mater Chem C. 2025;13(14):7307-7317.[DOI]
-
32. Sachan P, Mahapatra A, Sai Channapragada LA, Kaur R, Sahay S, Chandra Mondal P. Electrosynthesis of molecular memory elements. Chem Sci. 2025;16(24):10990-11001.[DOI]
-
34. Mini H, Park JE, Jo A. The meniscus in SECCM: Formation, stability, and its impact on quantitative electrochemistry. J Electrochem Sci Technol. 2026;17(3):304-320.[DOI]
-
36. Wang YQ, Wang D. Electrochemical scanning tunneling microscopy in electrocatalysis. Curr Opin Electrochem. 2024;46:101512.[DOI]
-
38. Cherevan AS, Nandan SP, Roger I, Rongji L, Streb C, Eder D. Polyoxometalates on functional substrates: Concepts, synergies, and future perspectives. Adv Sci. 2020;7(8):1903511.[DOI]
-
41. Huez C, Guérin D, Lenfant S, Volatron F, Calame M, Perrin ML, et al. Redox-controlled conductance of polyoxometalate molecular junctions. Nanoscale. 2022;14(37):13790-13800.[DOI]
-
43. Vogelsberg E, Moors M, Sorokina AS, Ryndyk DA, Schmitz S, Freitag JS, et al. Solution-processed formation of DNA-origami-supported polyoxometalate multi-level switches with countercation-controlled conductance tunability. Chem Mater. 2023;35(14):5447-5457.[DOI]
-
44. Phark SH, Weber B, Yoshida Y, Forrester PR, Elbertse RJG, Stroscio JA, et al. Roadmap on atomically-engineered quantum platforms. Nano Futur. 2025;9(3):032001.[DOI]
-
45. Choi DJ, Phark SH, Heinrich AJ, Lorente N. Electron spin resonance with scanning tunneling microscopy: A tool for an on-surface quantum platform of identical qubits. Nanoscale Adv. 2025;7(15):4551-4558.[DOI]
-
47. Bechtel HA, Muller EA, Olmon RL, Martin MC, Raschke MB. Ultrabroadband infrared nanospectroscopic imaging. Proc Natl Acad Sci U S A. 2014;111(20):7191-7196.[DOI]
-
48. Vicente RA, Raju SP, Gomes HVN, Neckel IT, Tolentino HCN, Fernández PS. Development of electrochemical cells and their application for spatially resolved analysis using a multitechnique approach: From conventional experiments to X-ray nanoprobe beamlines. Anal Chem. 2023;95(44):16144-16152.
-
49. Mino L, Borfecchia E, Segura-Ruiz J, Giannini C, Martinez-Criado G, Lamberti C. Materials characterization by synchrotron X-ray microprobes and nanoprobes. Rev Mod Phys. 2018;90(2):025007.[DOI]
-
50. Vicente RA, Neckel IT, Fernández PS. Mastering the use of nanoprobes beamlines for electrochemistry: The importance of tracking radiation damage and exploring material heterogeneity. ACS Electrochem. 2025;1(3):286-293.[DOI]
-
51. Hermann RJ, Gordon MJ. Nanoscale optical microscopy and spectroscopy using near-field probes. Annu Rev Chem Biomol Eng. 2018;9:365-387.[DOI]
-
52. Willke P, Bilgeri T, Zhang X, Wang Y, Wolf C, Aubin H, et al. Coherent Spin control of single molecules on a surface. ACS Nano. 2021;15(11):17959-17965.[DOI]
-
53. Sellies L, Repp J. Electron spin resonance at the single-molecule scale. Angew Chem Int Ed. 2025;64(37):e202506539.[DOI]
-
54. Wiens R, Findlay CR, Baldwin SG, Kreplak L, Lee JM, Veres SP, et al. High spatial resolution (1.1 μm and 20 nm) FTIR polarization contrast imaging reveals pre-rupture disorder in damaged tendon. Faraday Discuss. 2016;187:555-573.[DOI]
-
55. Volatron F, Izzet G, Vuillaume D, Proust A. Unveiling polyoxometalate redox properties at the nanoscale. Comptes Rendus Chim. 2024;27(G1):255-268.[DOI]
-
56. Chen JJ, Vilà-Nadal L, Solé-Daura A, Chisholm G, Minato T, Busche C, et al. Effective storage of electrons in water by the formation of highly reduced polyoxometalate clusters. J Am Chem Soc. 2022;144(20):8951-8960.[DOI]
-
57. Guillén Soler M, Rauschen R, Laws K, Sharma A, Grocholski N, McNulty M, et al. ElectroChemputer with integrated monitoring for programmable electrochemistry. Chem. 2026;12(3):102907.[DOI]
-
58. Ai F, Lu YC. Coordination chemistry in advanced redox-active electrolyte designs. Nat Rev Mater. 2025;10(12):929-946.[DOI]
-
59. Hörner G, Weber B. Switching spin-states: Spin crossover vs. Coordination-induced spin-state switching. Eur J Inorg Chem. 2024;27(35):e202400471.[DOI]
-
60. Saygili Y, Stojanovic M, Flores-Díaz N, Zakeeruddin SM, Vlachopoulos N, Grätzel M, et al. Metal coordination complexes as redox mediators in regenerative dye-sensitized solar cells. Inorganics. 2019;7(3):30.[DOI]
-
62. Bertini I, Luchinat C, Parigi G, Ravera E. Transition metal ions: Shift and relaxation. In: Solution NMR of paramagnetic molecules. Amsterdam: Elsevier; 2017. p. 175-253.[DOI]
-
64. Zhang Y, Torres-Cavanillas R, Yan X, Zeng Y, Jiang M, Clemente-León M, et al. Spin crossover iron complexes with spin transition near room temperature based on nitrogen ligands containing aromatic rings: From molecular design to functional devices. Chem Soc Rev. 2024;53(17):8764-8789.
-
66. Das A, Ren Y, Hessin C, Desage-El Murr M. Copper catalysis with redox-active ligands. Beilstein J Org Chem. 2020;16:858-870.[DOI]
-
69. Tahoun M, Gee CT, McCoy VE, Sander PM, Müller CE. Chemistry of porphyrins in fossil plants and animals. RSC Adv. 2021;11(13):7552-7563.[DOI]
-
73. Gray HB, Winkler JR. Electron transfer in proteins. Annu Rev Biochem. 1996;65:537-561.[DOI]
-
74. Cojocariu I, Carlotto S, Zamborlini G, Jugovac M, Schio L, Floreano L, et al. Reversible redox reactions in metal-supported porphyrin: The role of spin and oxidation state. J Mater Chem C. 2021;9(37):12559-12565.[DOI]
-
75. Zhang Y, Si W, Hou X, Gao Q, Zhao C, Liang R, et al. Single-molecule spin devices: Fundamentals, advances, and prospects. Nano Res. 2026;19(6):94908552.[DOI]
-
77. Anjass M, Lowe GA, Streb C. Molecular vanadium oxides for energy conversion and energy storage: Current trends and emerging opportunities. Angew Chem Int Ed. 2021;60(14):7522-7532.[DOI]
-
78. Arya N, Philipp T, Greiner S, Steiner M, Kranz C, Anjass M. Reversible electrodeposition of potassium-bridged molecular vanadium oxides: A new approach towards multi-electron storage. Angew Chem Int Ed. 2023;62(35):e202306170.[DOI]
Copyright
© The Author(s) 2027. This is an Open Access article licensed under a Creative Commons Attribution 4.0 International License (https://creativecommons.org/licenses/by/4.0/), which permits unrestricted use, sharing, adaptation, distribution and reproduction in any medium or format, for any purpose, even commercially, as long as you give appropriate credit to the original author(s) and the source, provide a link to the Creative Commons license, and indicate if changes were made.
Publisher’s Note
Science Exploration remains a neutral stance on jurisdictional claims in published
maps
and institutional affiliations. The views expressed in this article are solely those
of
the author(s) and do not reflect the opinions of the Editors or the publisher.
Share And Cite


Science Exploration Style
Al-Shamery N, Vicente RA, Zocher C, Kersting B, Monakhov K. Bridging the scales: Advanced scanning probe microscopy across molecular redox regimes. Smart Mater Devices. 2027;3:202646. https://doi.org/10.70401/smd.2026.0054
Tips
Copy completed.
Submit a Manuscript
Author Instructions
Cite this Article
Article Metrics
0
View
0
Download
Cited
Article Updates

- Abstract
- Keywords
- 1. Introduction
- 2. Why Redox-Active Molecular Systems Require a Regime-Aware Framework
- 3. Bulk Ensemble Regime
- 4. Confined Nanoscale Ensembles
- 5. Single-Molecule and Few-Molecule Regimes: EC-STM, STM/STS, and ESR-STM
- 6. Advanced Optical and Synchrotron-Coupled SPM
- 7. Toward Regime-Bridging Molecular Analysis
- 8. From POMs to Related Ionic and Redox-Active Molecular Materials
- 9. Conclusion
- Acknowledgments
- Authors contribution
- Conflicts of interest
- Ethical approval
- Consent to participate
- Consent for publication
- Availability of data and materials
- Funding
- References
- Copyright
 Get citation
Science Exploration Style

Al-Shamery N, Vicente RA, Zocher C, Kersting B, Monakhov K. Bridging the scales: Advanced scanning probe microscopy across molecular redox regimes. Smart Mater Devices. 2027;3:202646. https://doi.org/10.70401/smd.2026.0054
copy
Share Link
copy