Dynamic kinetic resolution (DKR) of configurationally labile biaryls via enantioselective C−H functionalization

Jie Zhang
#
,
Xuanzhu Huo
#
,
Yidan Liu
#
,
Xiu-Feng Hou

Can Zhu
*
*Correspondence to:
Can Zhu, Research Center for Molecular Recognition and Synthesis, State Key Laboratory of Green Chemical Synthesis and Conversion, Department of Chemistry, Fudan University, Shanghai 200438, China.
E-mail: zhucan@fudan.edu.cn
Chiral Chem. 2027;3:202627. 10.70401/cc.2026.0036
Received: June 11, 2026Accepted: August 06, 2026Published: August 07, 2026
Abstract
Owing to their stable axial chirality, atropisomeric biaryl compounds have found widespread applications in medicinal chemistry, natural product chemistry, and agricultural chemistry. Up to now, two principal strategies have been established for the synthesis of axially chiral biaryls: the enantioselective construction of the aryl-aryl axis via asymmetric coupling reactions, and the stereoselective modification of a pre-existing axis, which includes asymmetric construction of one aromatic ring, desymmetrization of prochiral substrates, and dynamic kinetic resolution (DKR) of biaryl skeletons. Among these strategies, enantioselective C−H functionalization avoids the use of pre-functionalized substrates and installs sterically demanding groups that enhance rotational barriers, thereby serving as an ideal DKR tool for the construction of atropisomeric biaryls directly from biaryl substrates with low rotational barriers. In this review, based on the nature of the directing groups, we summarize three distinct strategies for enantioselective C−H functionalization of biaryls with free axial rotation: directing group (DG)-mediated, transient DG-enabled, and DG-free approaches. The mechanisms and substrate scopes of selected representative reactions are also briefly discussed.
Graphical Abstract
Keywords
Dynamic kinetic resolution, atropisomeric biaryls, axial rotation, C−H functionalization, directing group
1. Introduction
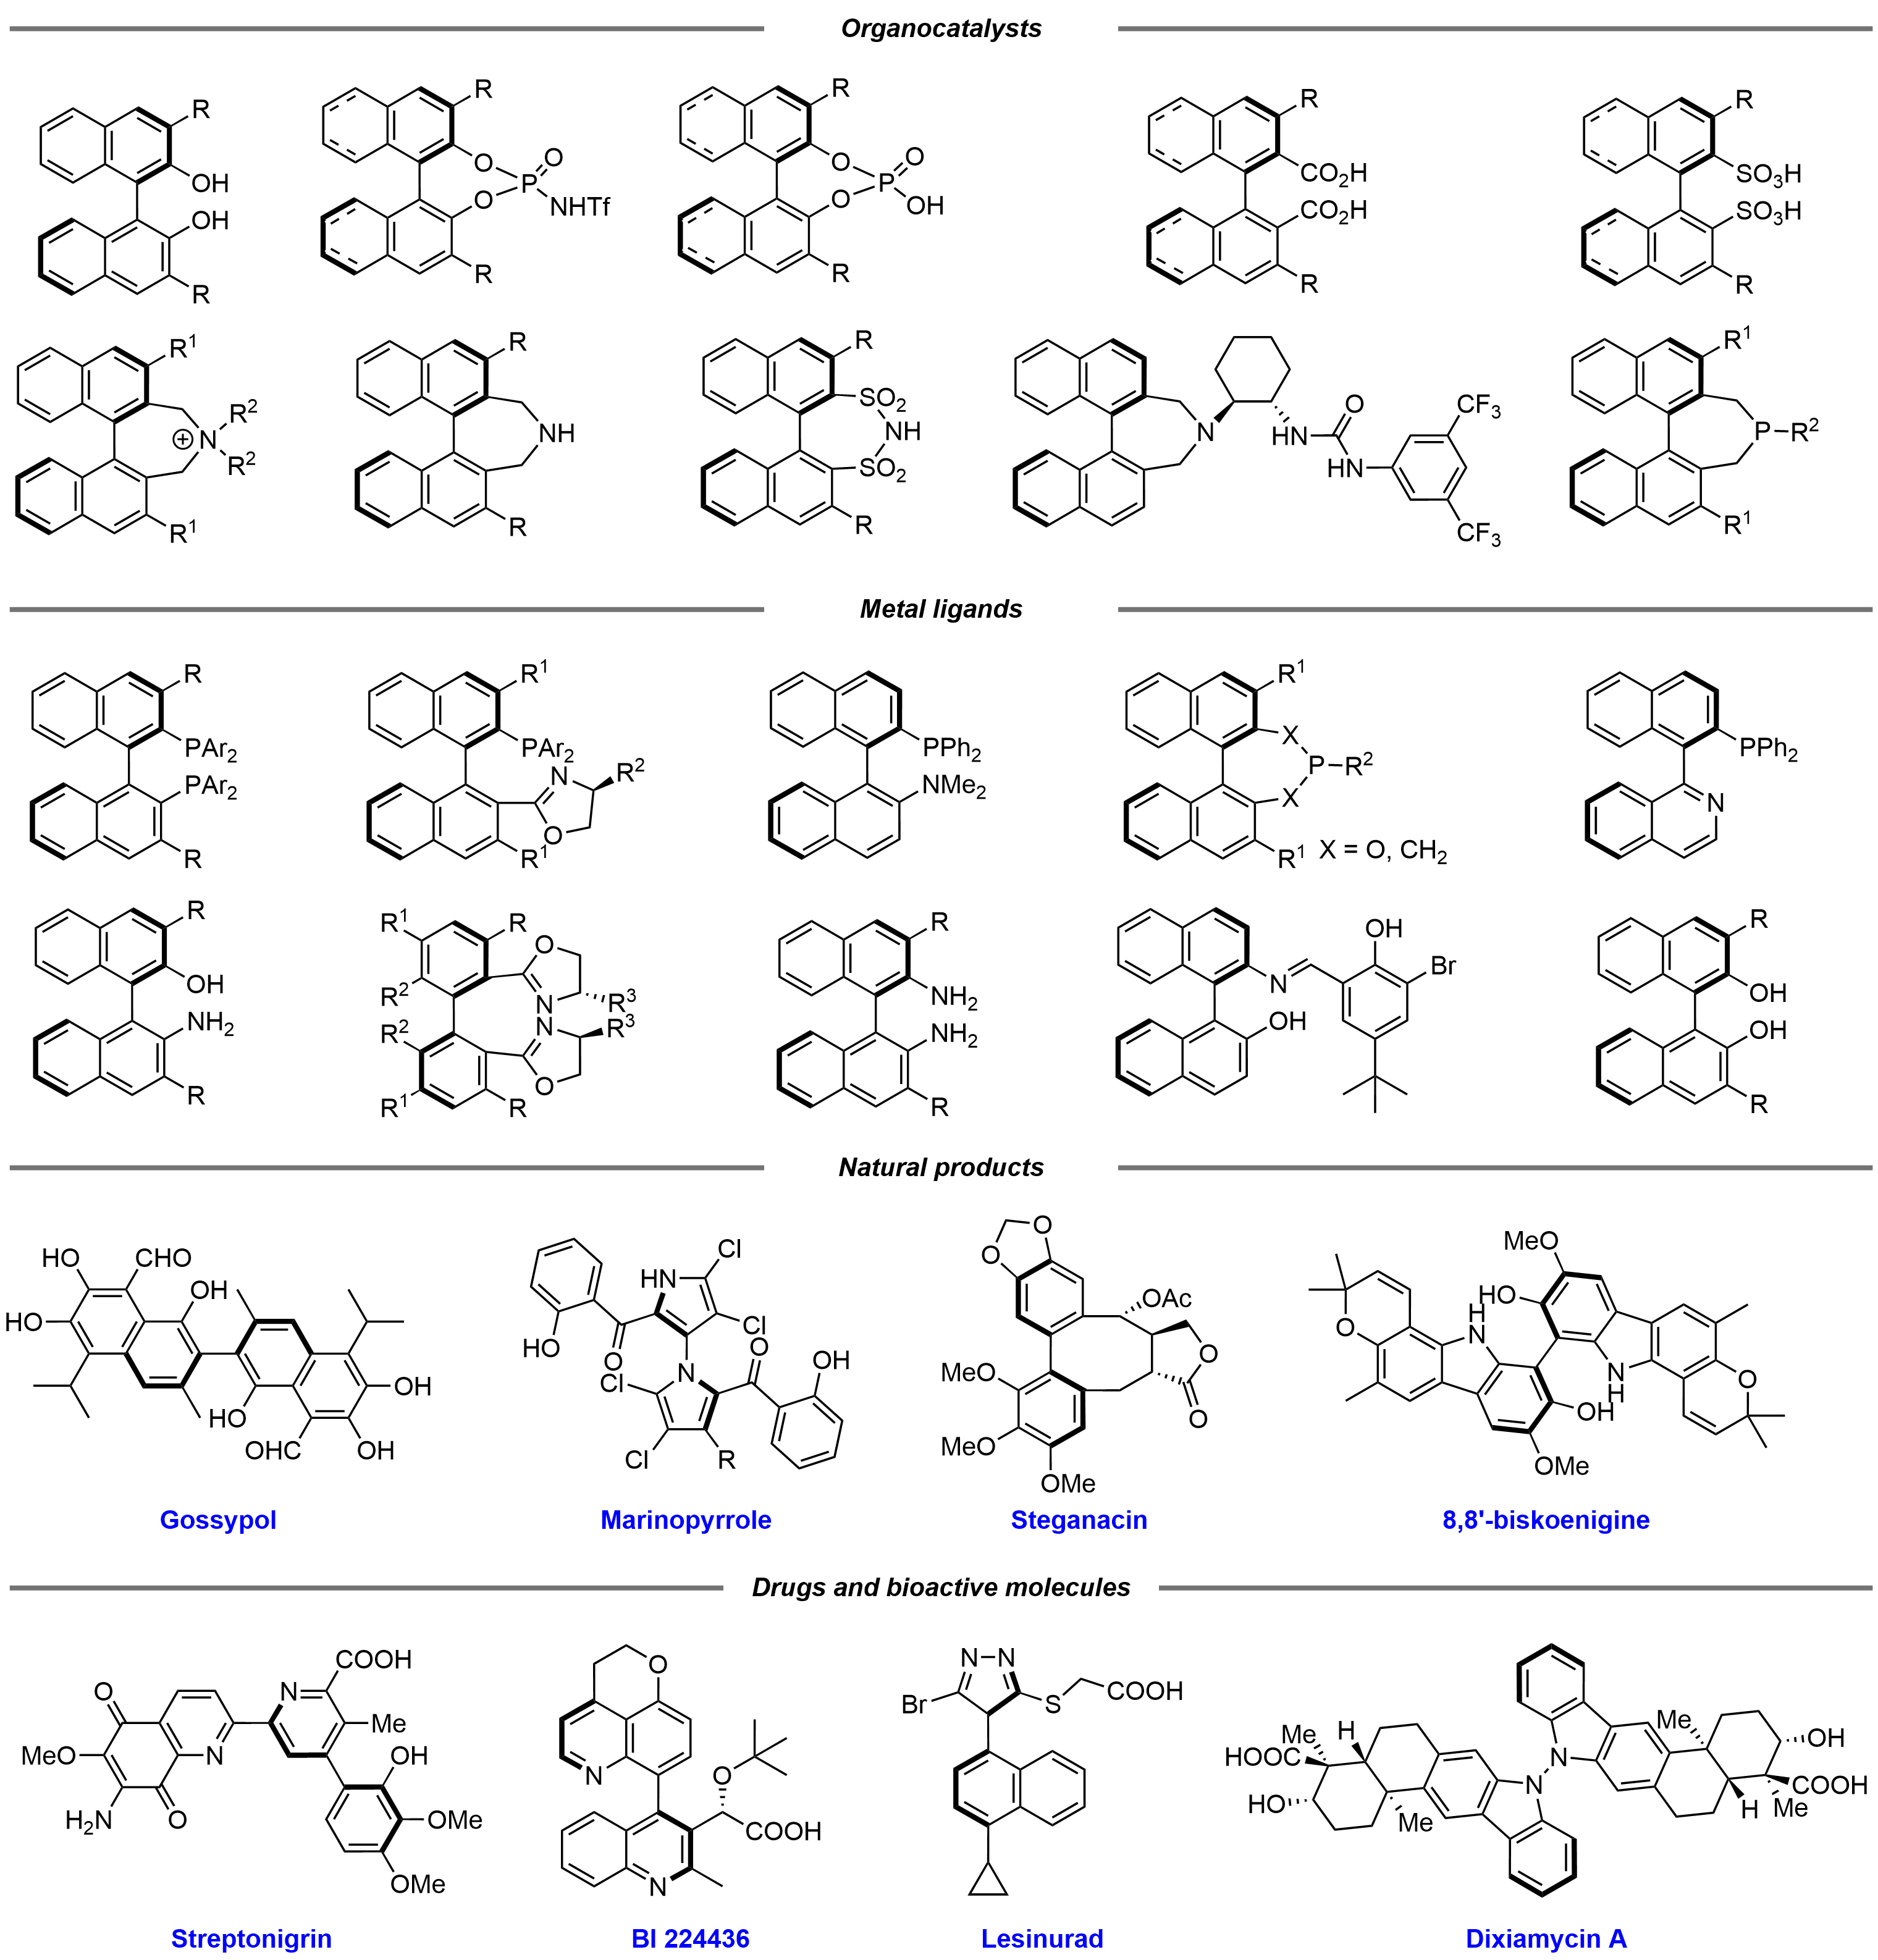
Axial chirality constitutes a distinctive type of stereoisomerism, where the chiral environment originates not from a conventional stereogenic center but from restricted rotation around a single bond, thereby generating an inherent chiral axis[1-3]. This stereochemical phenomenon is most prominently manifested in biaryl frameworks, in which steric substituents significantly elevate the rotational barrier and thus furnish configurationally stable stereoisomers termed atropisomers[4-6]. Axially chiral skeletons are recognized as privileged structural motifs, which are ubiquitously distributed in natural products[7] and bioactive molecules[8]. Meanwhile, they also serve as the core building blocks of chiral ligands and organocatalysts widely employed in asymmetric catalytic synthesis (Scheme 1)[9,10]. In this context, the development of efficient and highly stereoselective synthetic strategies for constructing such axially chiral architectures remains a vibrant and fundamentally important research frontier in organic chemistry.
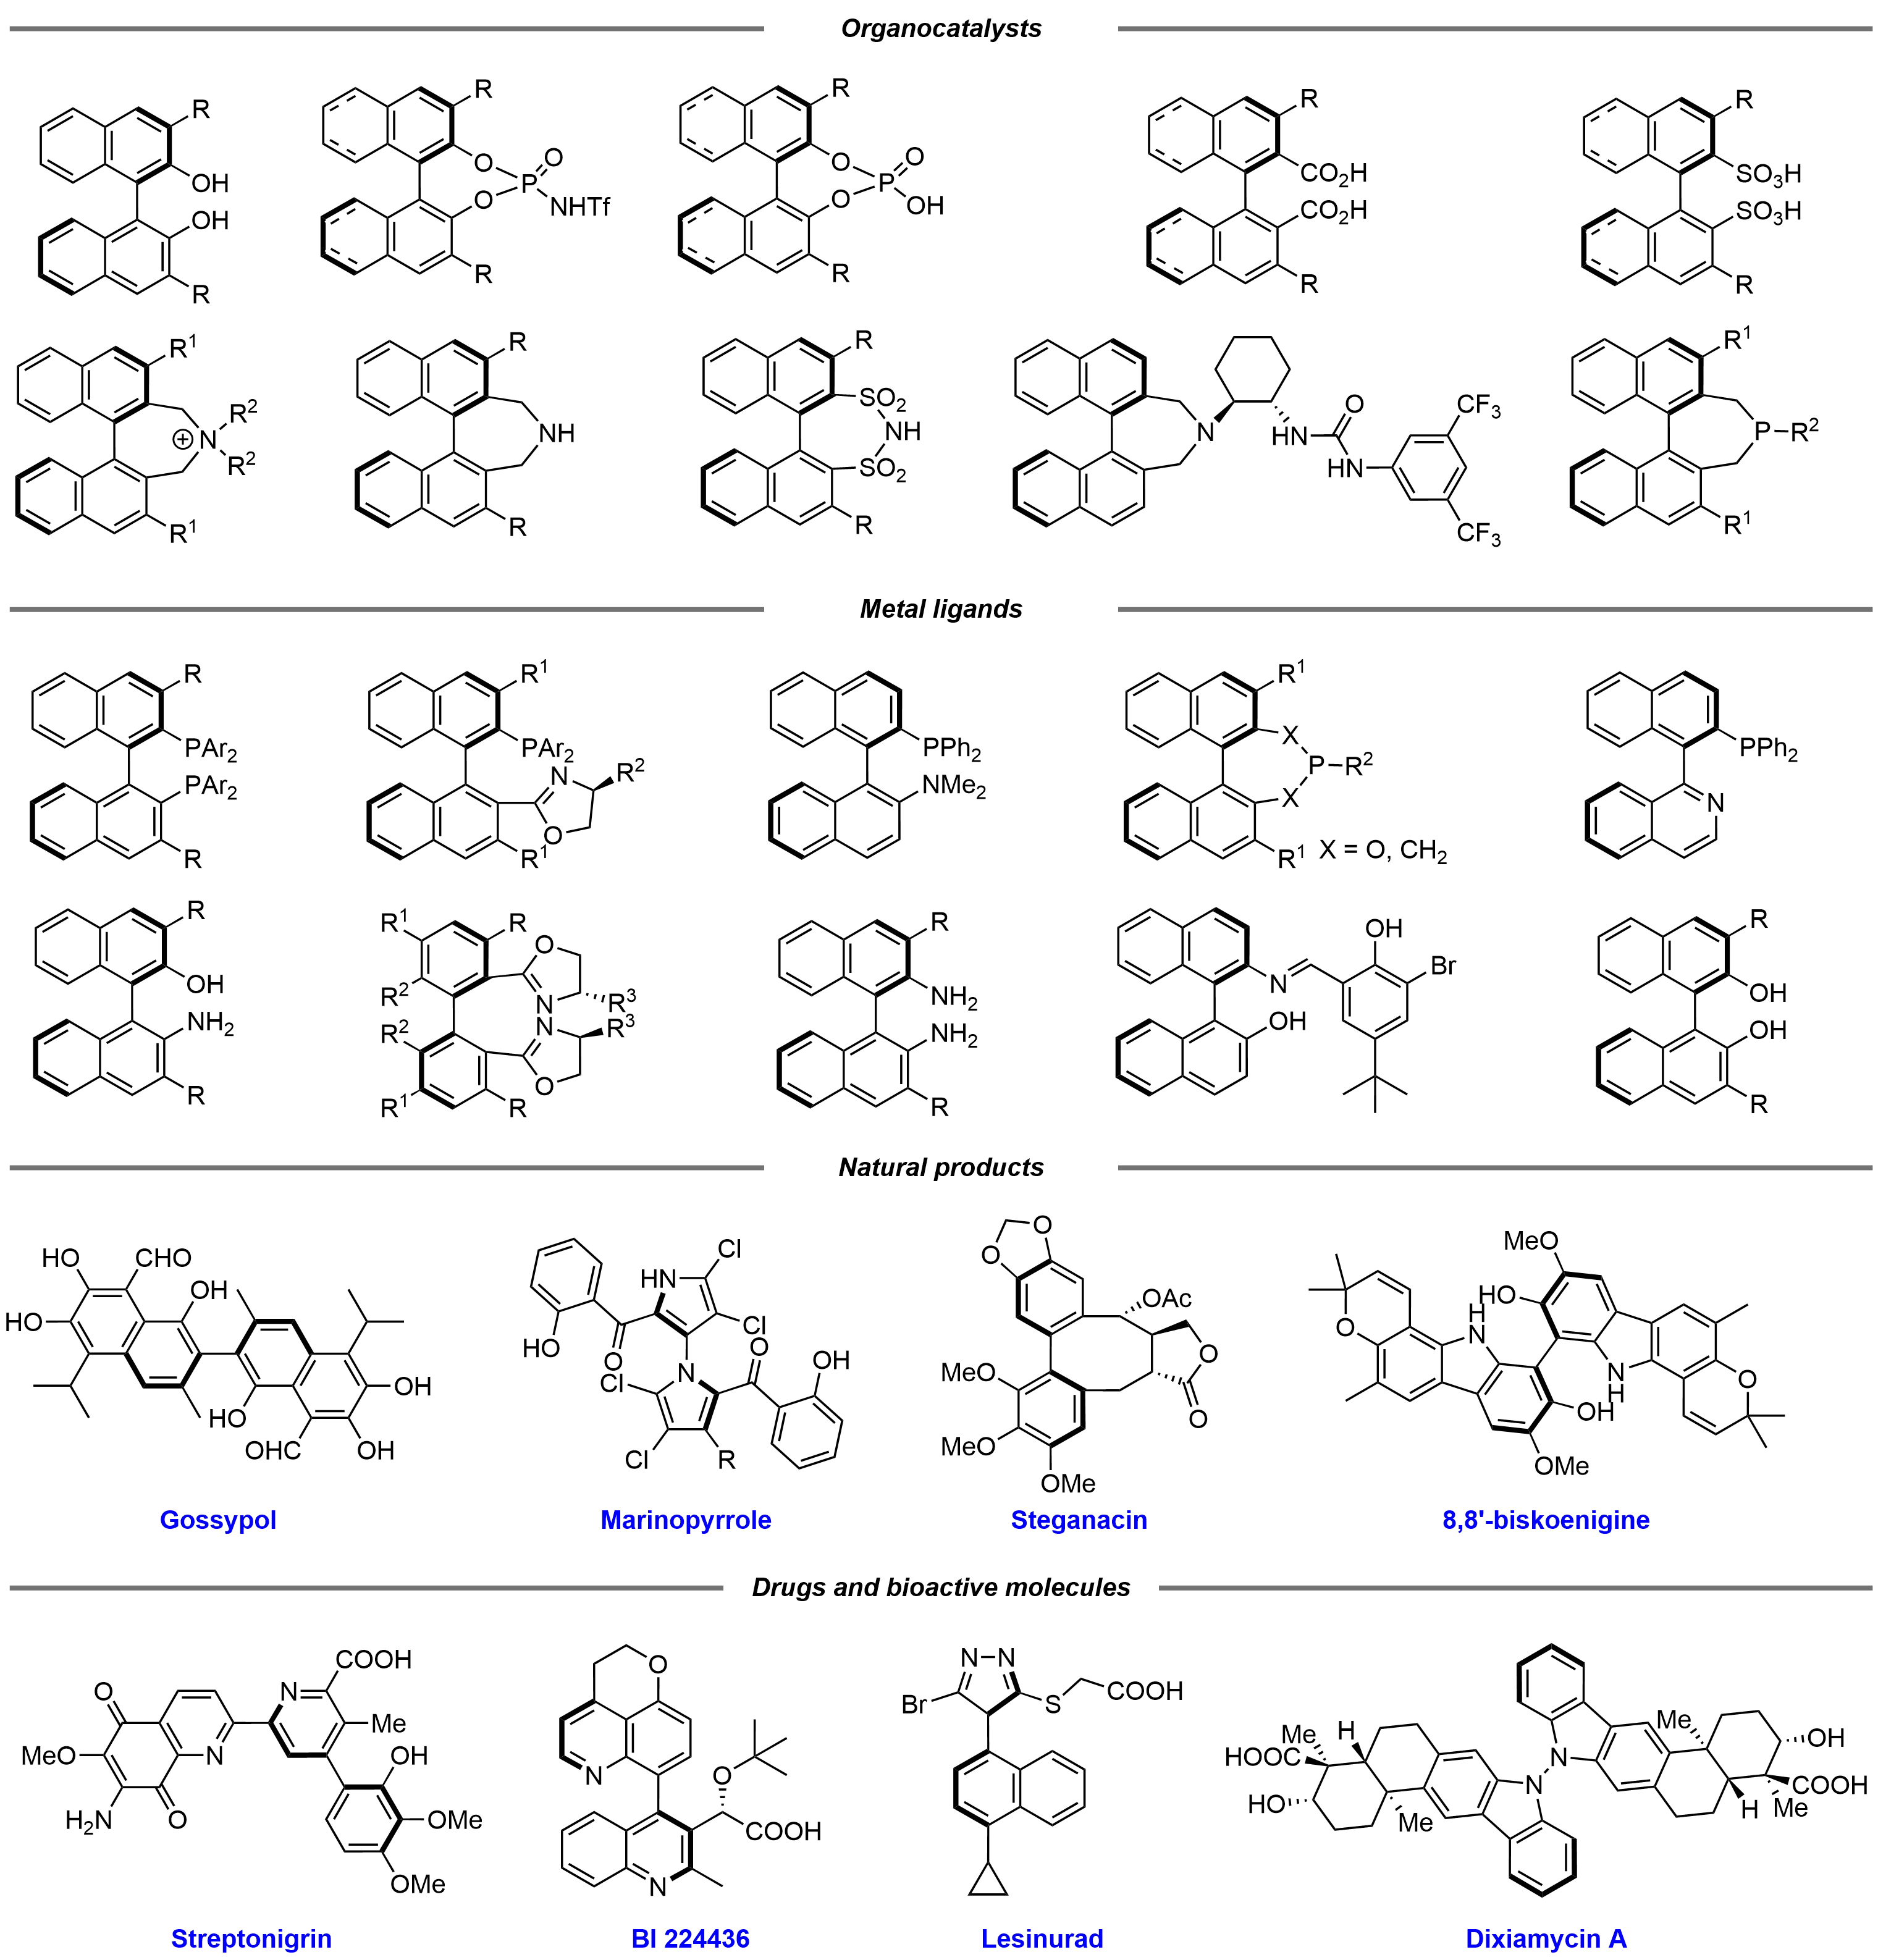
Scheme 1. Selected examples of atropisomeric biaryls as chiral ligands, organocatalysts, and natural products.
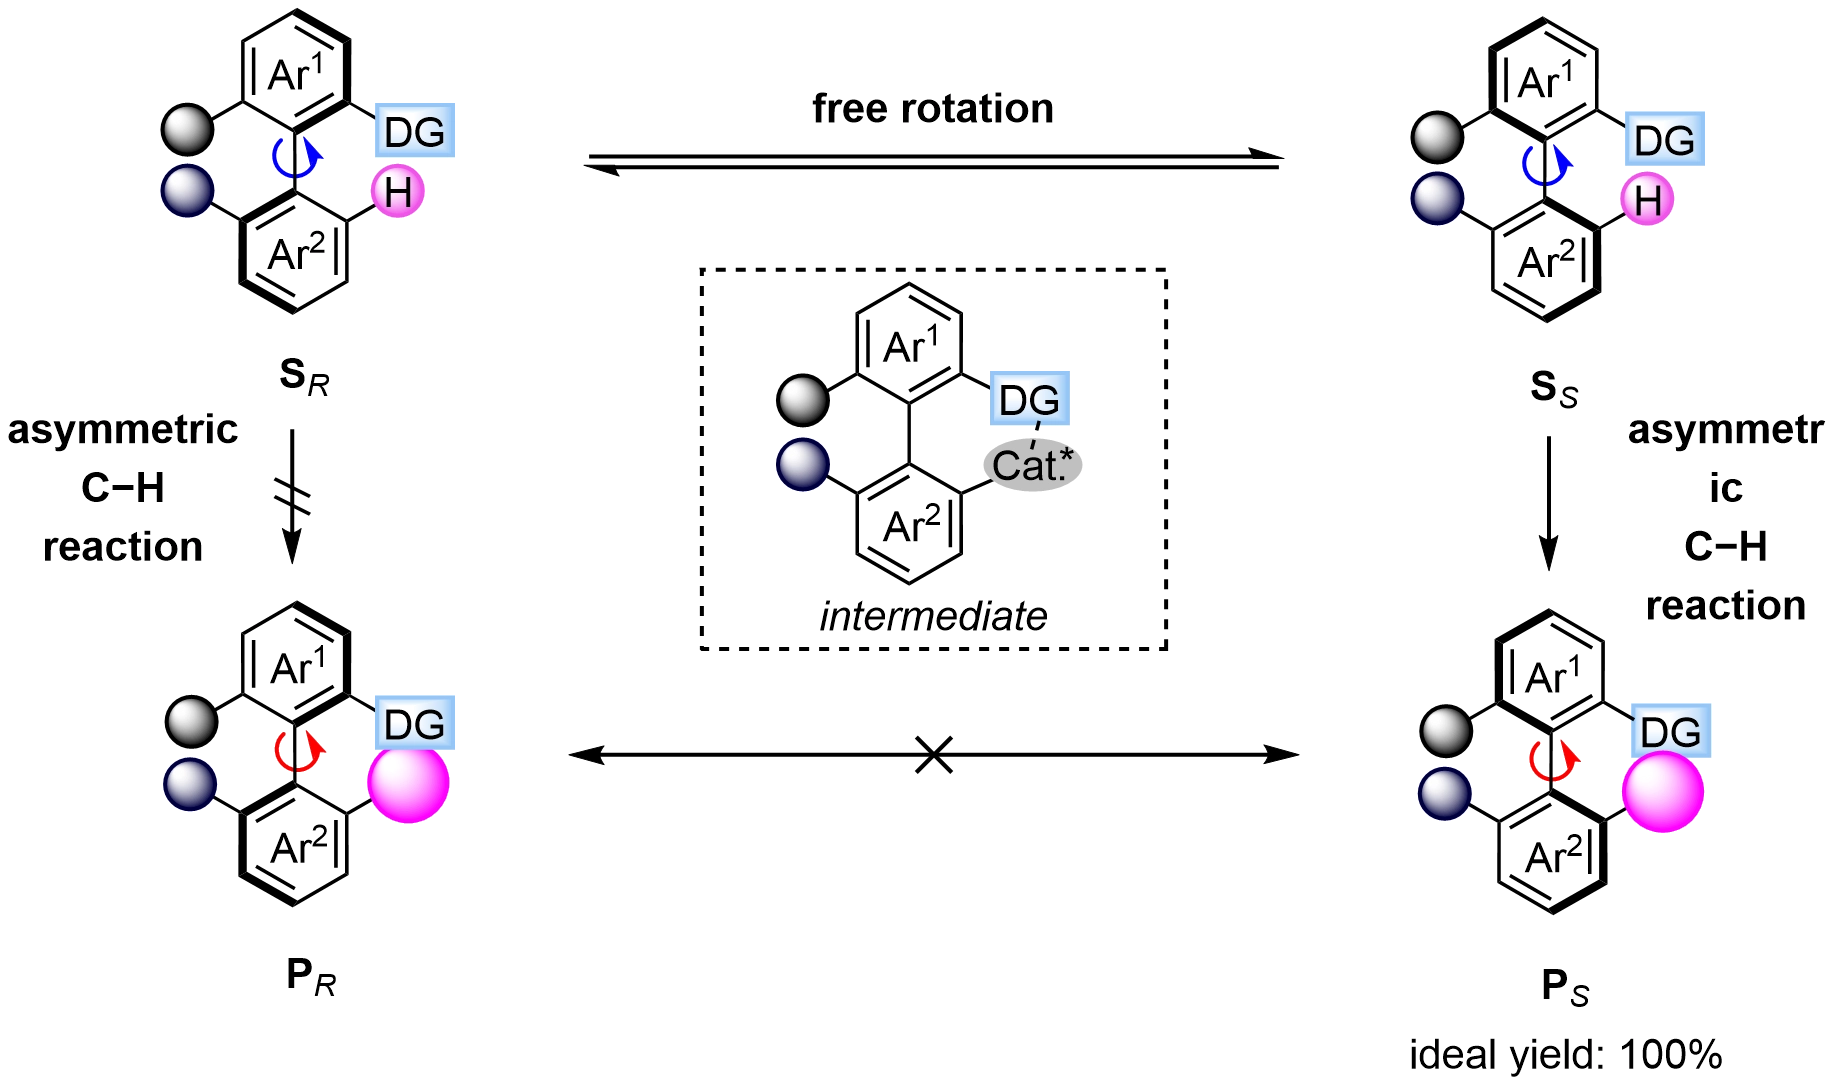
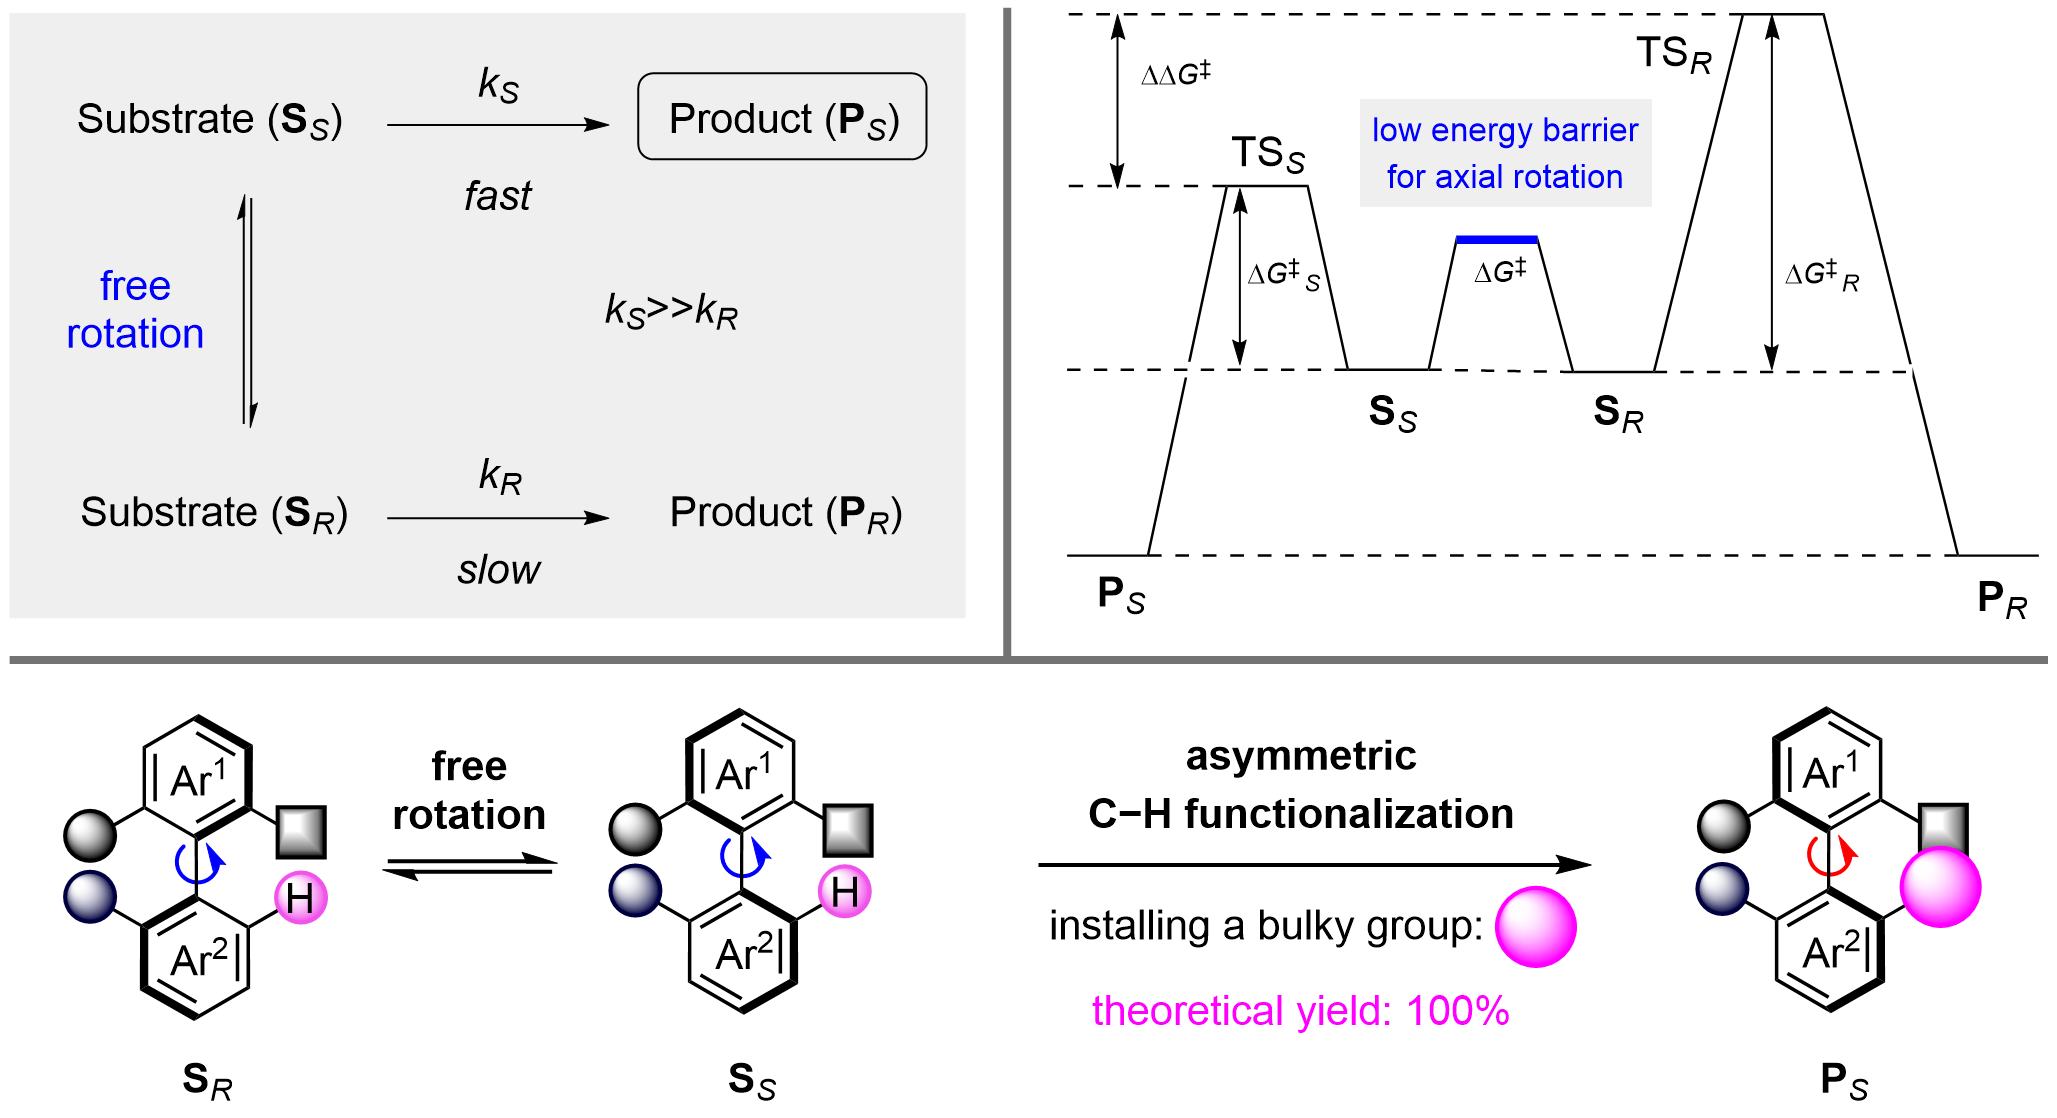
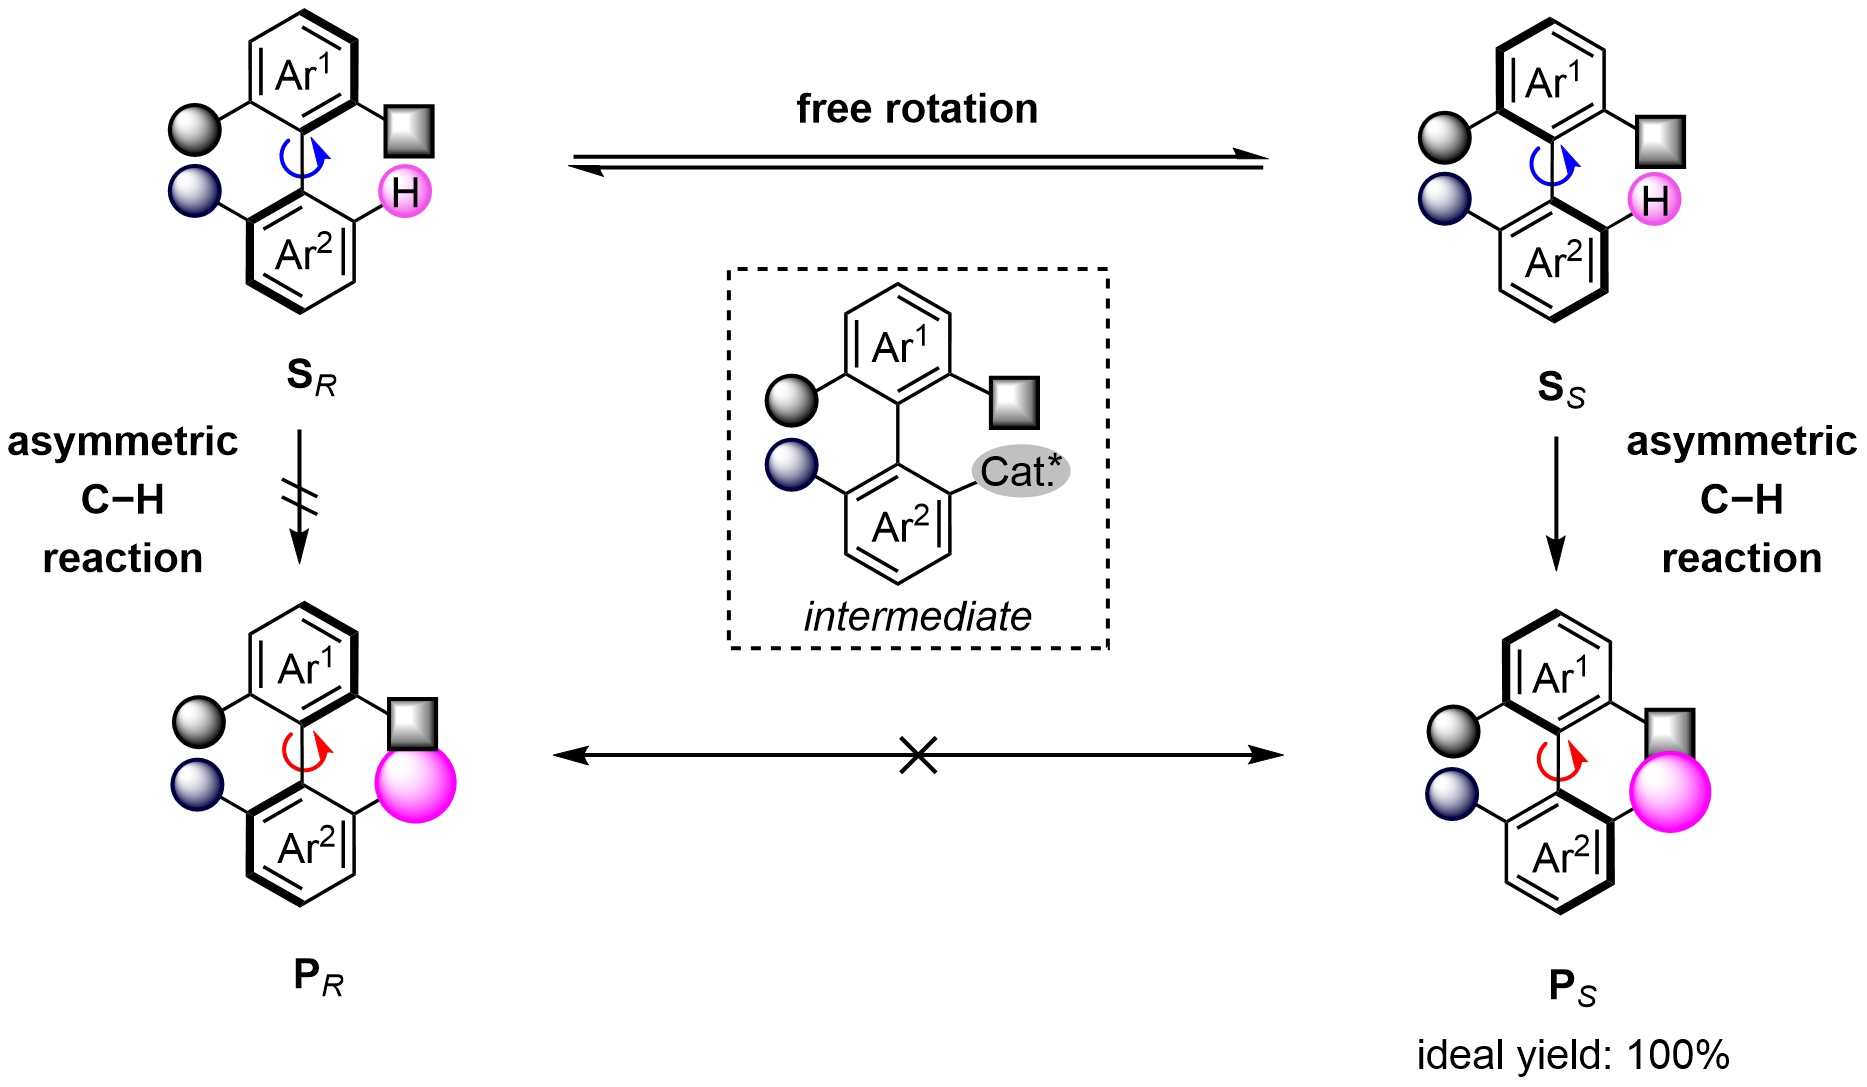
Kinetic resolution (KR) represents a fundamental strategy in asymmetric synthesis. In this process, the two enantiomers of a racemic substrate undergo transformation at distinctly different reaction rates (kS>>kR), allowing access to enantiomerically enriched compounds[11-13]. However, conventional KR is inherently limited by a maximum theoretical yield of 50%[14,15]. In contrast, dynamic kinetic resolution (DKR) circumvents this bottleneck by combining rapid in situ racemization with enantioselective functionalization, offering a highly atom- and step-economical pathway to access axially chiral frameworks (Scheme 2)[16-19].
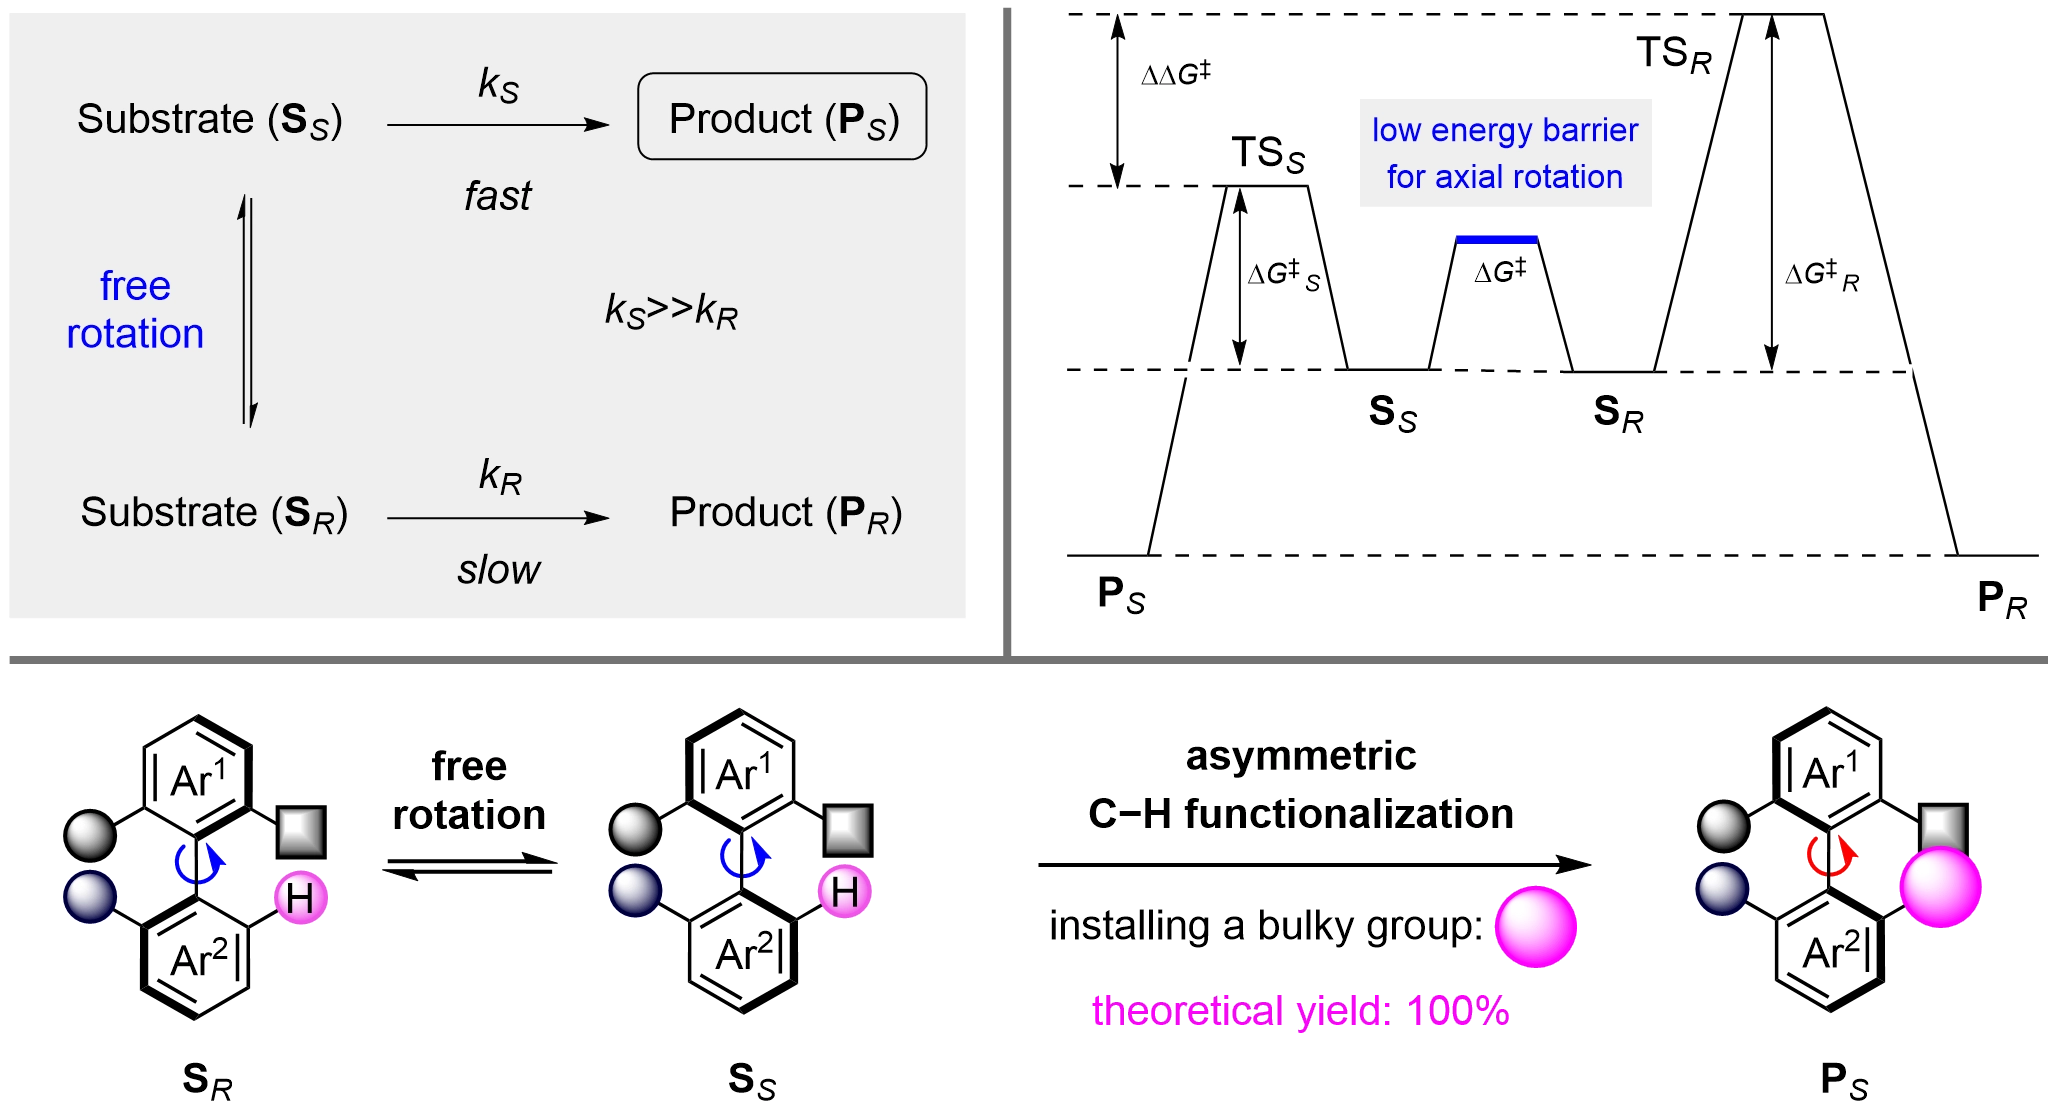
Scheme 2. DKR of biaryls with free axial rotation. DKR: dynamic kinetic resolution.
Integrating C−H functionalization with DKR allows the direct catalytic construction of stereogenic axes from simple precursors. In this cooperative process, C−H activation plays an indispensable role by installing sterically demanding substituents at the ortho-positions of biaryl scaffolds. This selectively restricts the free rotation of the aryl-aryl single bond, thereby locking the resulting molecule into a configurationally stable atropisomer. Importantly, while asymmetric catalysis on configurationally stable racemates typically yields conventional KR outcomes, employing configurationally labile biaryls as substrates allows C−H functionalization to achieve DKR. Under DKR conditions, the unreacted enantiomer undergoes continuous and reversible racemization, enabled by the low rotational barrier of the starting material prior to functionalization. For this transformation to proceed efficiently, the rate of substrate racemization must surpass that of the stereodetermining step of C−H functionalization, ensuring that both enantiomers are productively funneled toward a single enantiopure product.
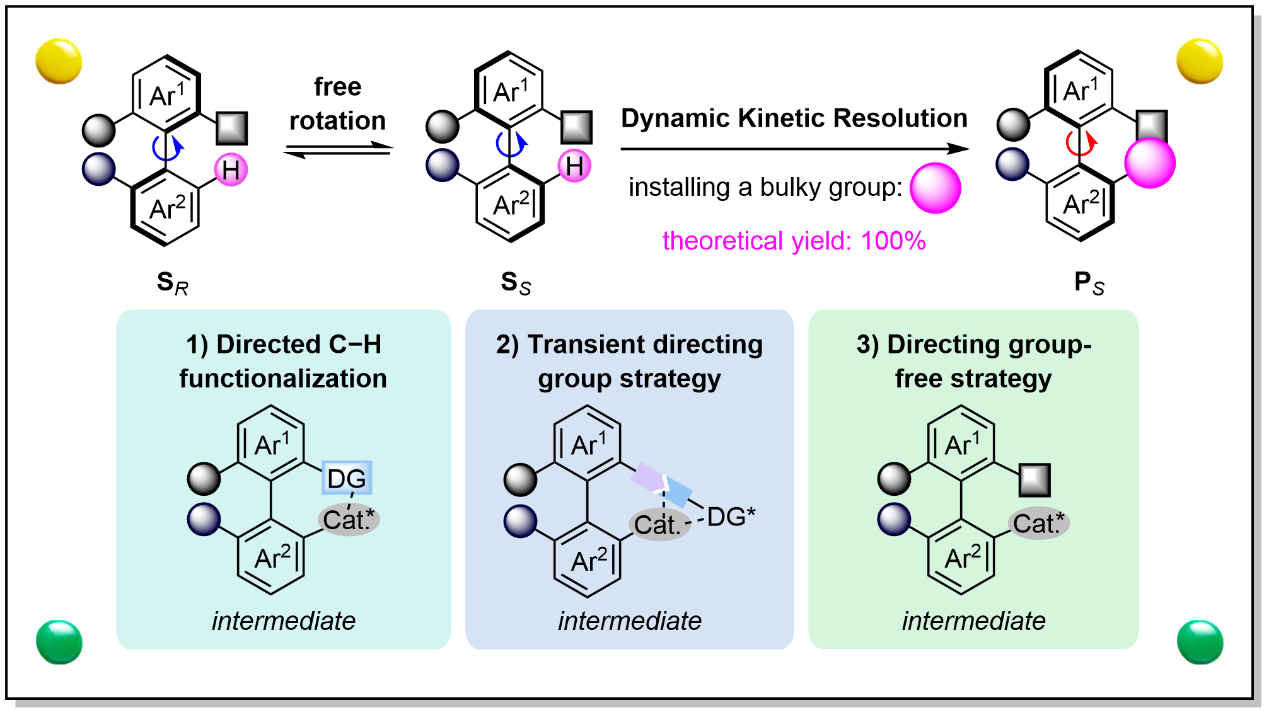
From the perspective of substrate design and directing strategies, three major synthetic approaches have been established within this framework: 1) Directed C−H activation strategy, which relies on a preinstalled directing group (DG) to achieve excellent regioselectivity and precise stereocontrol; 2) Transient directing group (TDG) strategy, which enables the in-situ generation of a directing template through reversible noncovalent interactions. This approach obviates the extra steps for permanent directing group installation and removal, while substantially broadening the substrate scope; 3) DG-free C−H functionalization strategy, which eliminates the need for either permanent or transient directing groups by incorporating heteroatoms (such as nitrogen or oxygen) into the aryl rings to modulate electronic properties, thereby leveraging noncovalent interactions and steric hindrance to achieve high enantioselectivity.
This review comprehensively summarizes the recent progress achieved across various research groups in the field of C−H functionalization-enabled DKR. It systematically collates and categorizes state-of-the-art advances toward the dynamic kinetic construction of biaryl axially chiral compounds via C−H activation strategies. We anticipate that this review will stimulate further in-depth investigations within this research area and open up new opportunities for the continued development and application of such synthetic methodologies.
2. DKR of Biaryls with Free Axial Rotations via C−H Functionalization
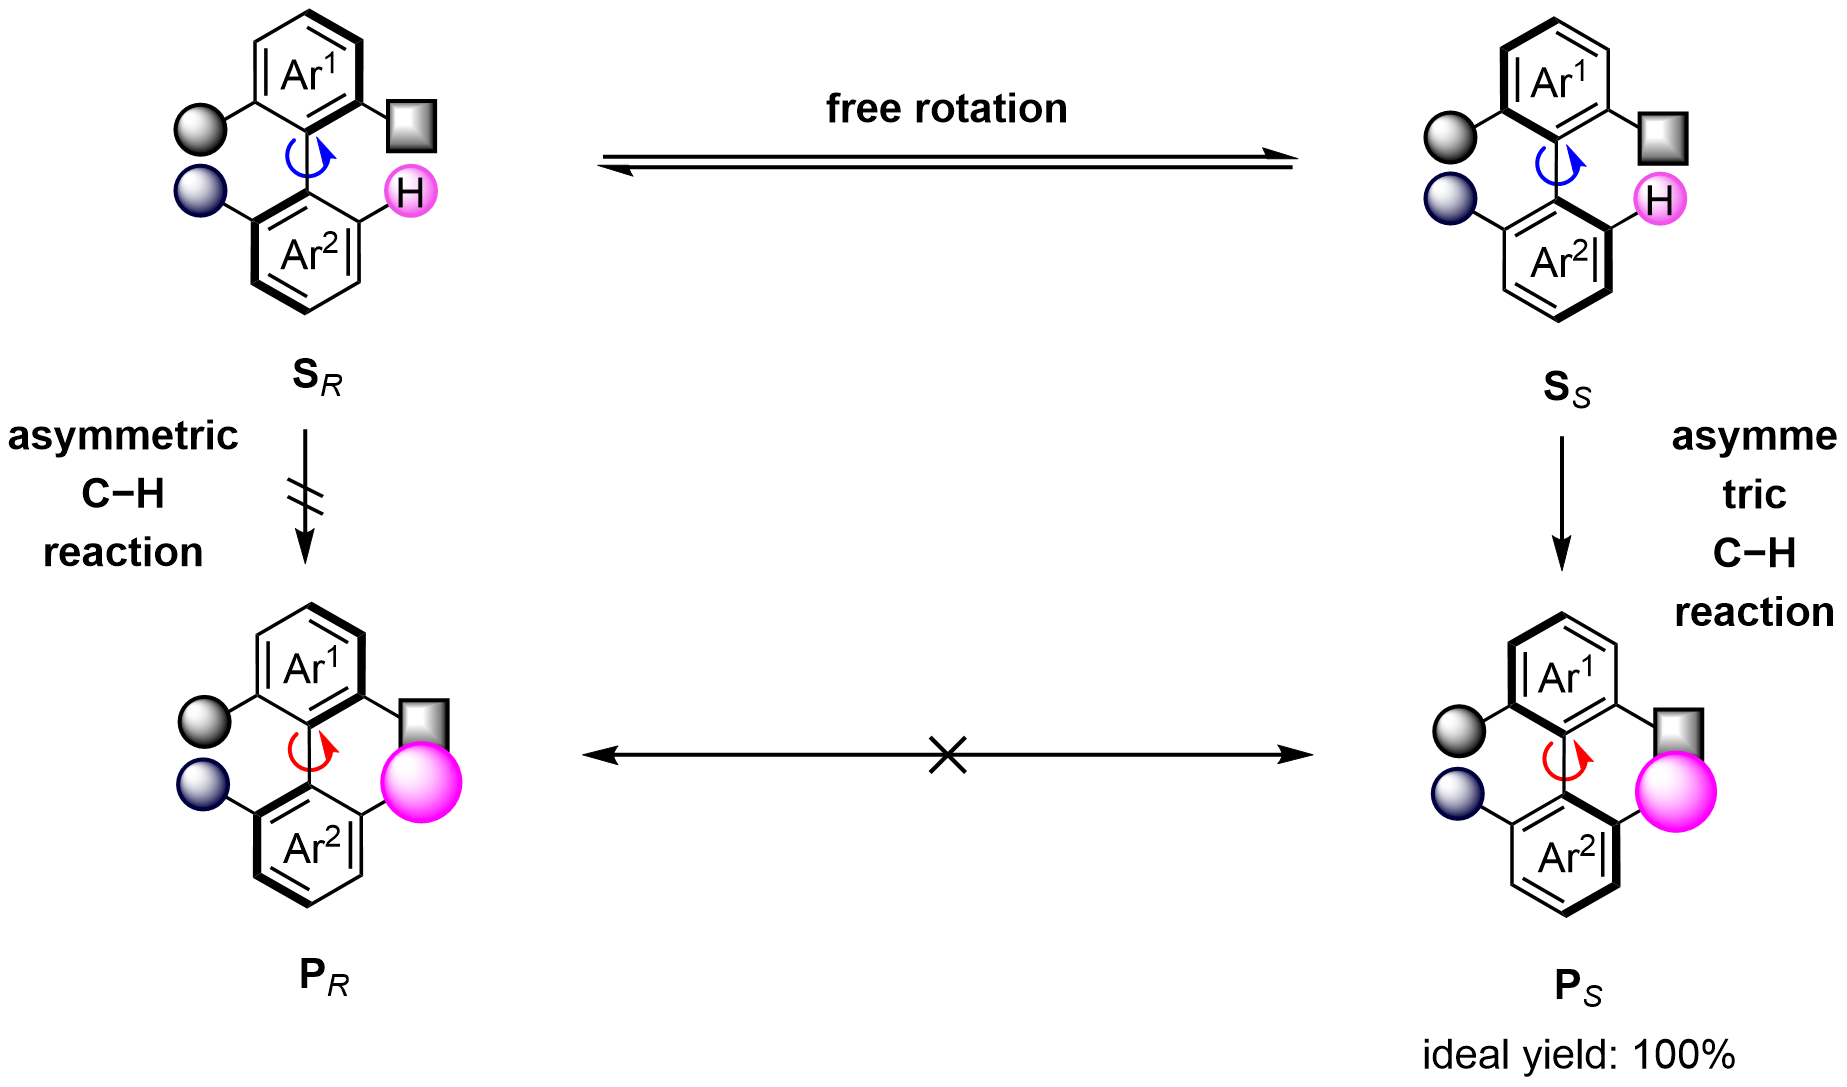
The group of hydrogen was generally considered as the smallest substituent at the neighboring positions of the axis in biaryl compounds, resulting in low rotational barriers and configurational instability. Asymmetric C−H functionalization occurs to these biaryl compounds, replacing hydrogen with a larger substituent to increase the rotational barrier. Therefore, direct C−H functionalization can be elegantly merged into the DKR of biaryls with free axial rotation. Enantioselective functionalization of a C−H bond can be realized via strategies including asymmetric metal catalysis or cooperative catalysis between transition metals and chiral ligands, as well as asymmetric addition or substitution reactions by organocatalysts, leading to the generation of stable hindered biaryl products (Scheme 3)[20-23].
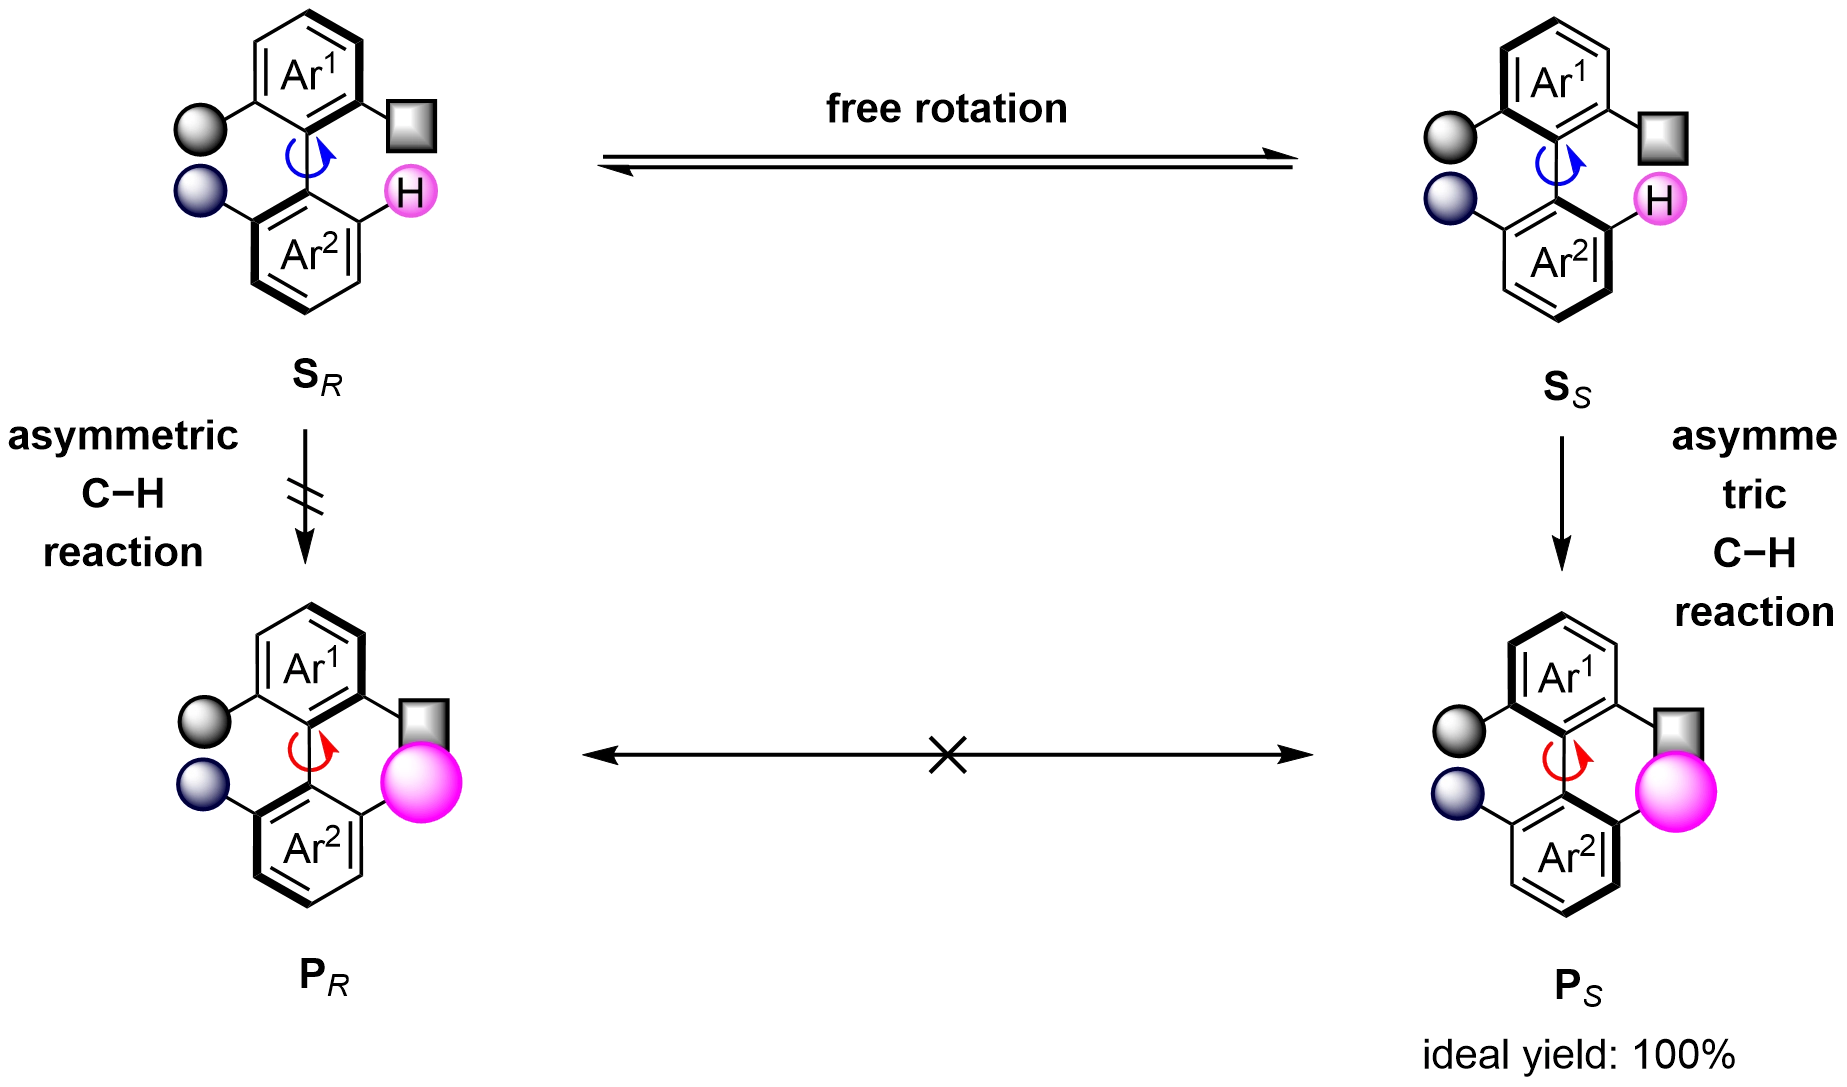
Scheme 3. DKR of biaryls with free axial rotation via C−H functionalization. DKR: dynamic kinetic resolution.
2.1 DKR of biaryls with free axial rotation via directed C−H functionalization
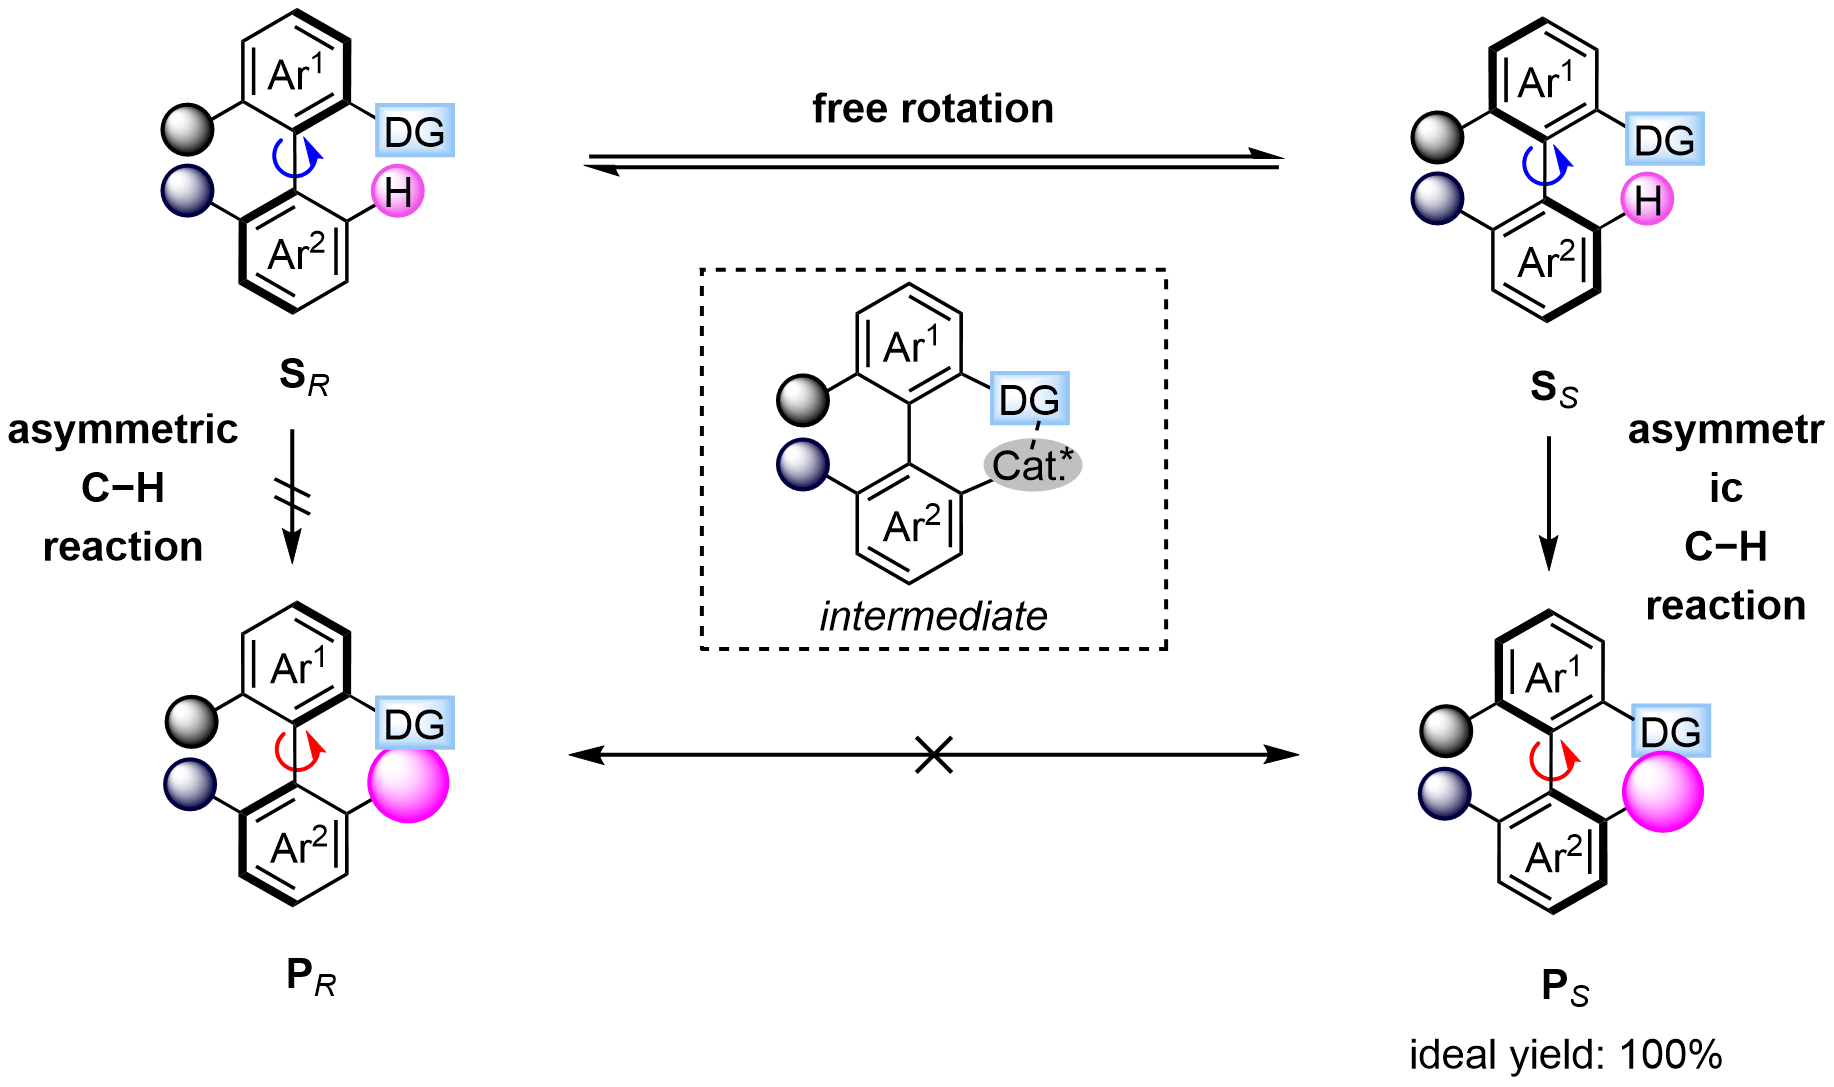
The introduction of a suitable directing group in biaryl skeletons can not only control the site-selectivity in the C−H functionalization process, but also play a crucial role in stabilizing the chiral axis by coordination with chiral metal catalysts to increase its steric hindrance. The groups of You, Shi, and many others have independently developed various catalytic systems to access axially chiral biaryls via the DKR strategy (Scheme 4).
Scheme 4. DKR of biaryls with free axial rotation via directed C−H functionalization. DKR: dynamic kinetic resolution.
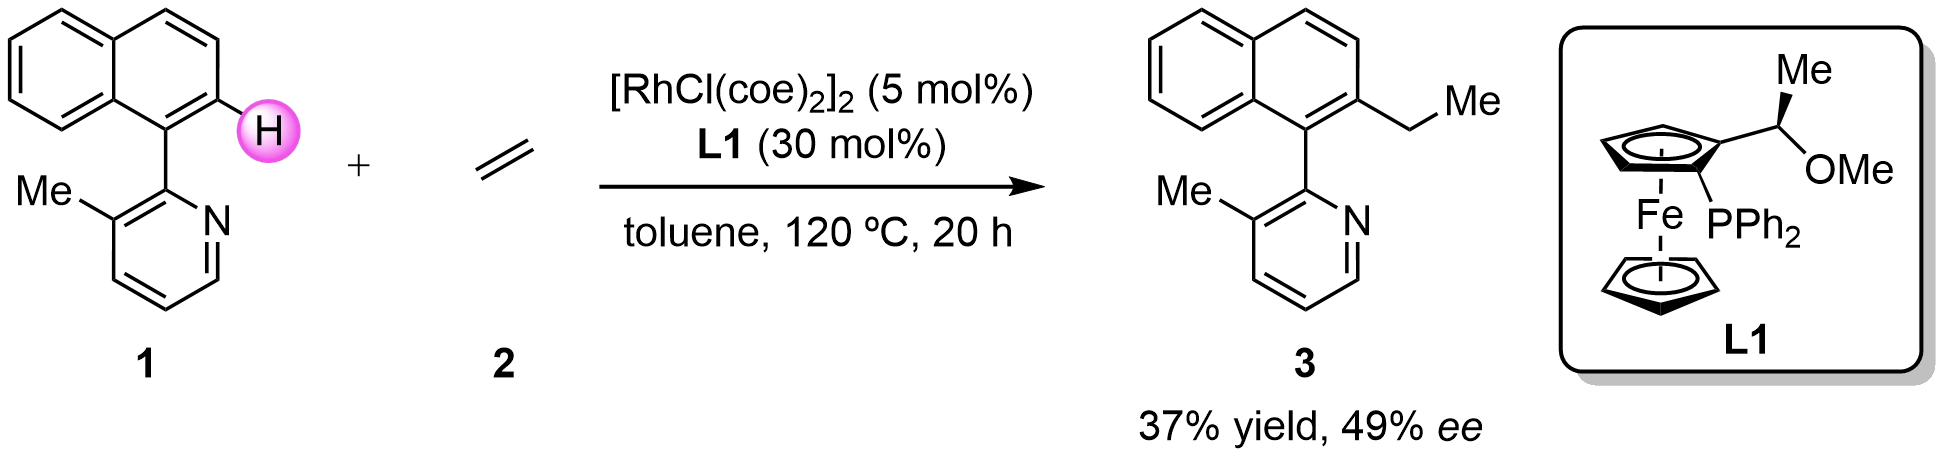
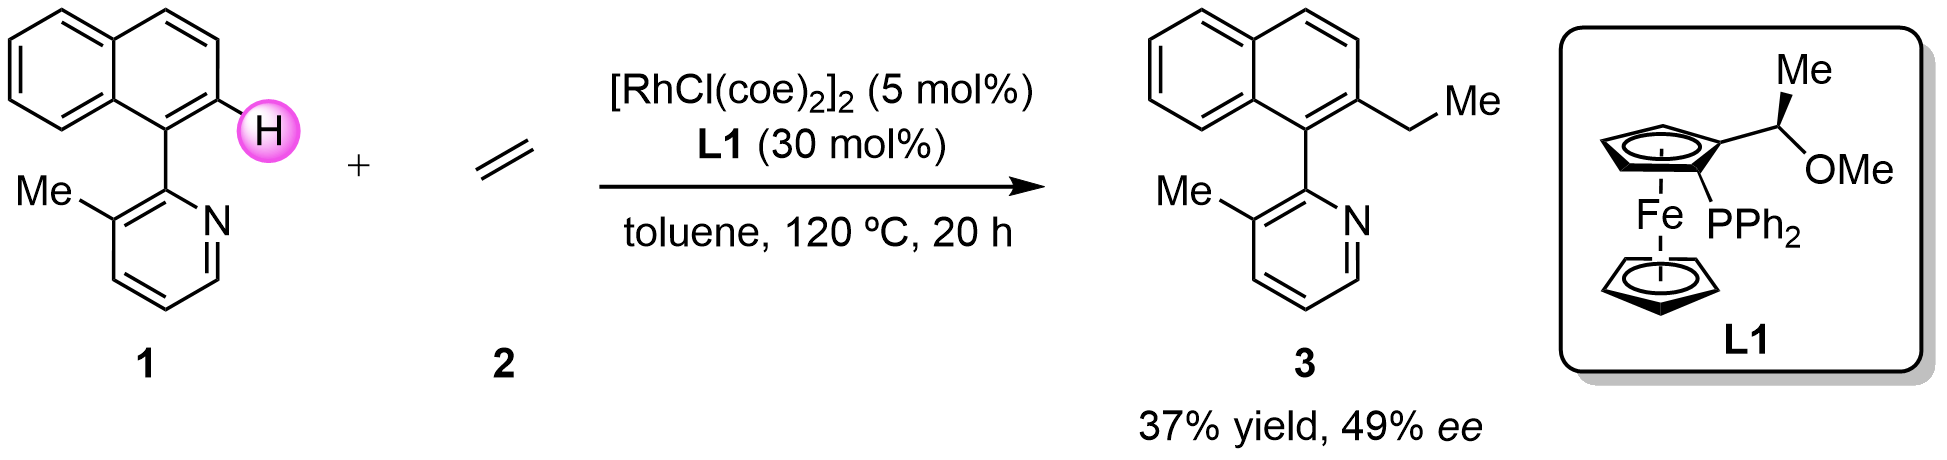
In 1972, the Pedersen group reported that 1-(1-naphthyl)isoquinoline possesses a low rotation barrier of only 18.6 kcal/mol. Due to this low barrier, the substrate readily undergoes in situ racemization at room temperature[24]. In 2000, the Murai group reported the first DKR of arylpyridines through the rhodium-catalyzed pyridine-directed asymmetric C−H alkylation with olefins (Scheme 5)[25]. [RhCl(coe)2]2 and L1 were used as catalysts to deliver products with alkylation occurring at the 2-position of the naphthalene ring, which prevented the axial rotation. Although the efficiency (37% yield) and enantioselectivity (49% ee) of the reaction are not satisfactory, this method establishes a fundamental basis for subsequent development of the DKR of biaryls via asymmetric C−H activation under transition metal catalysis.
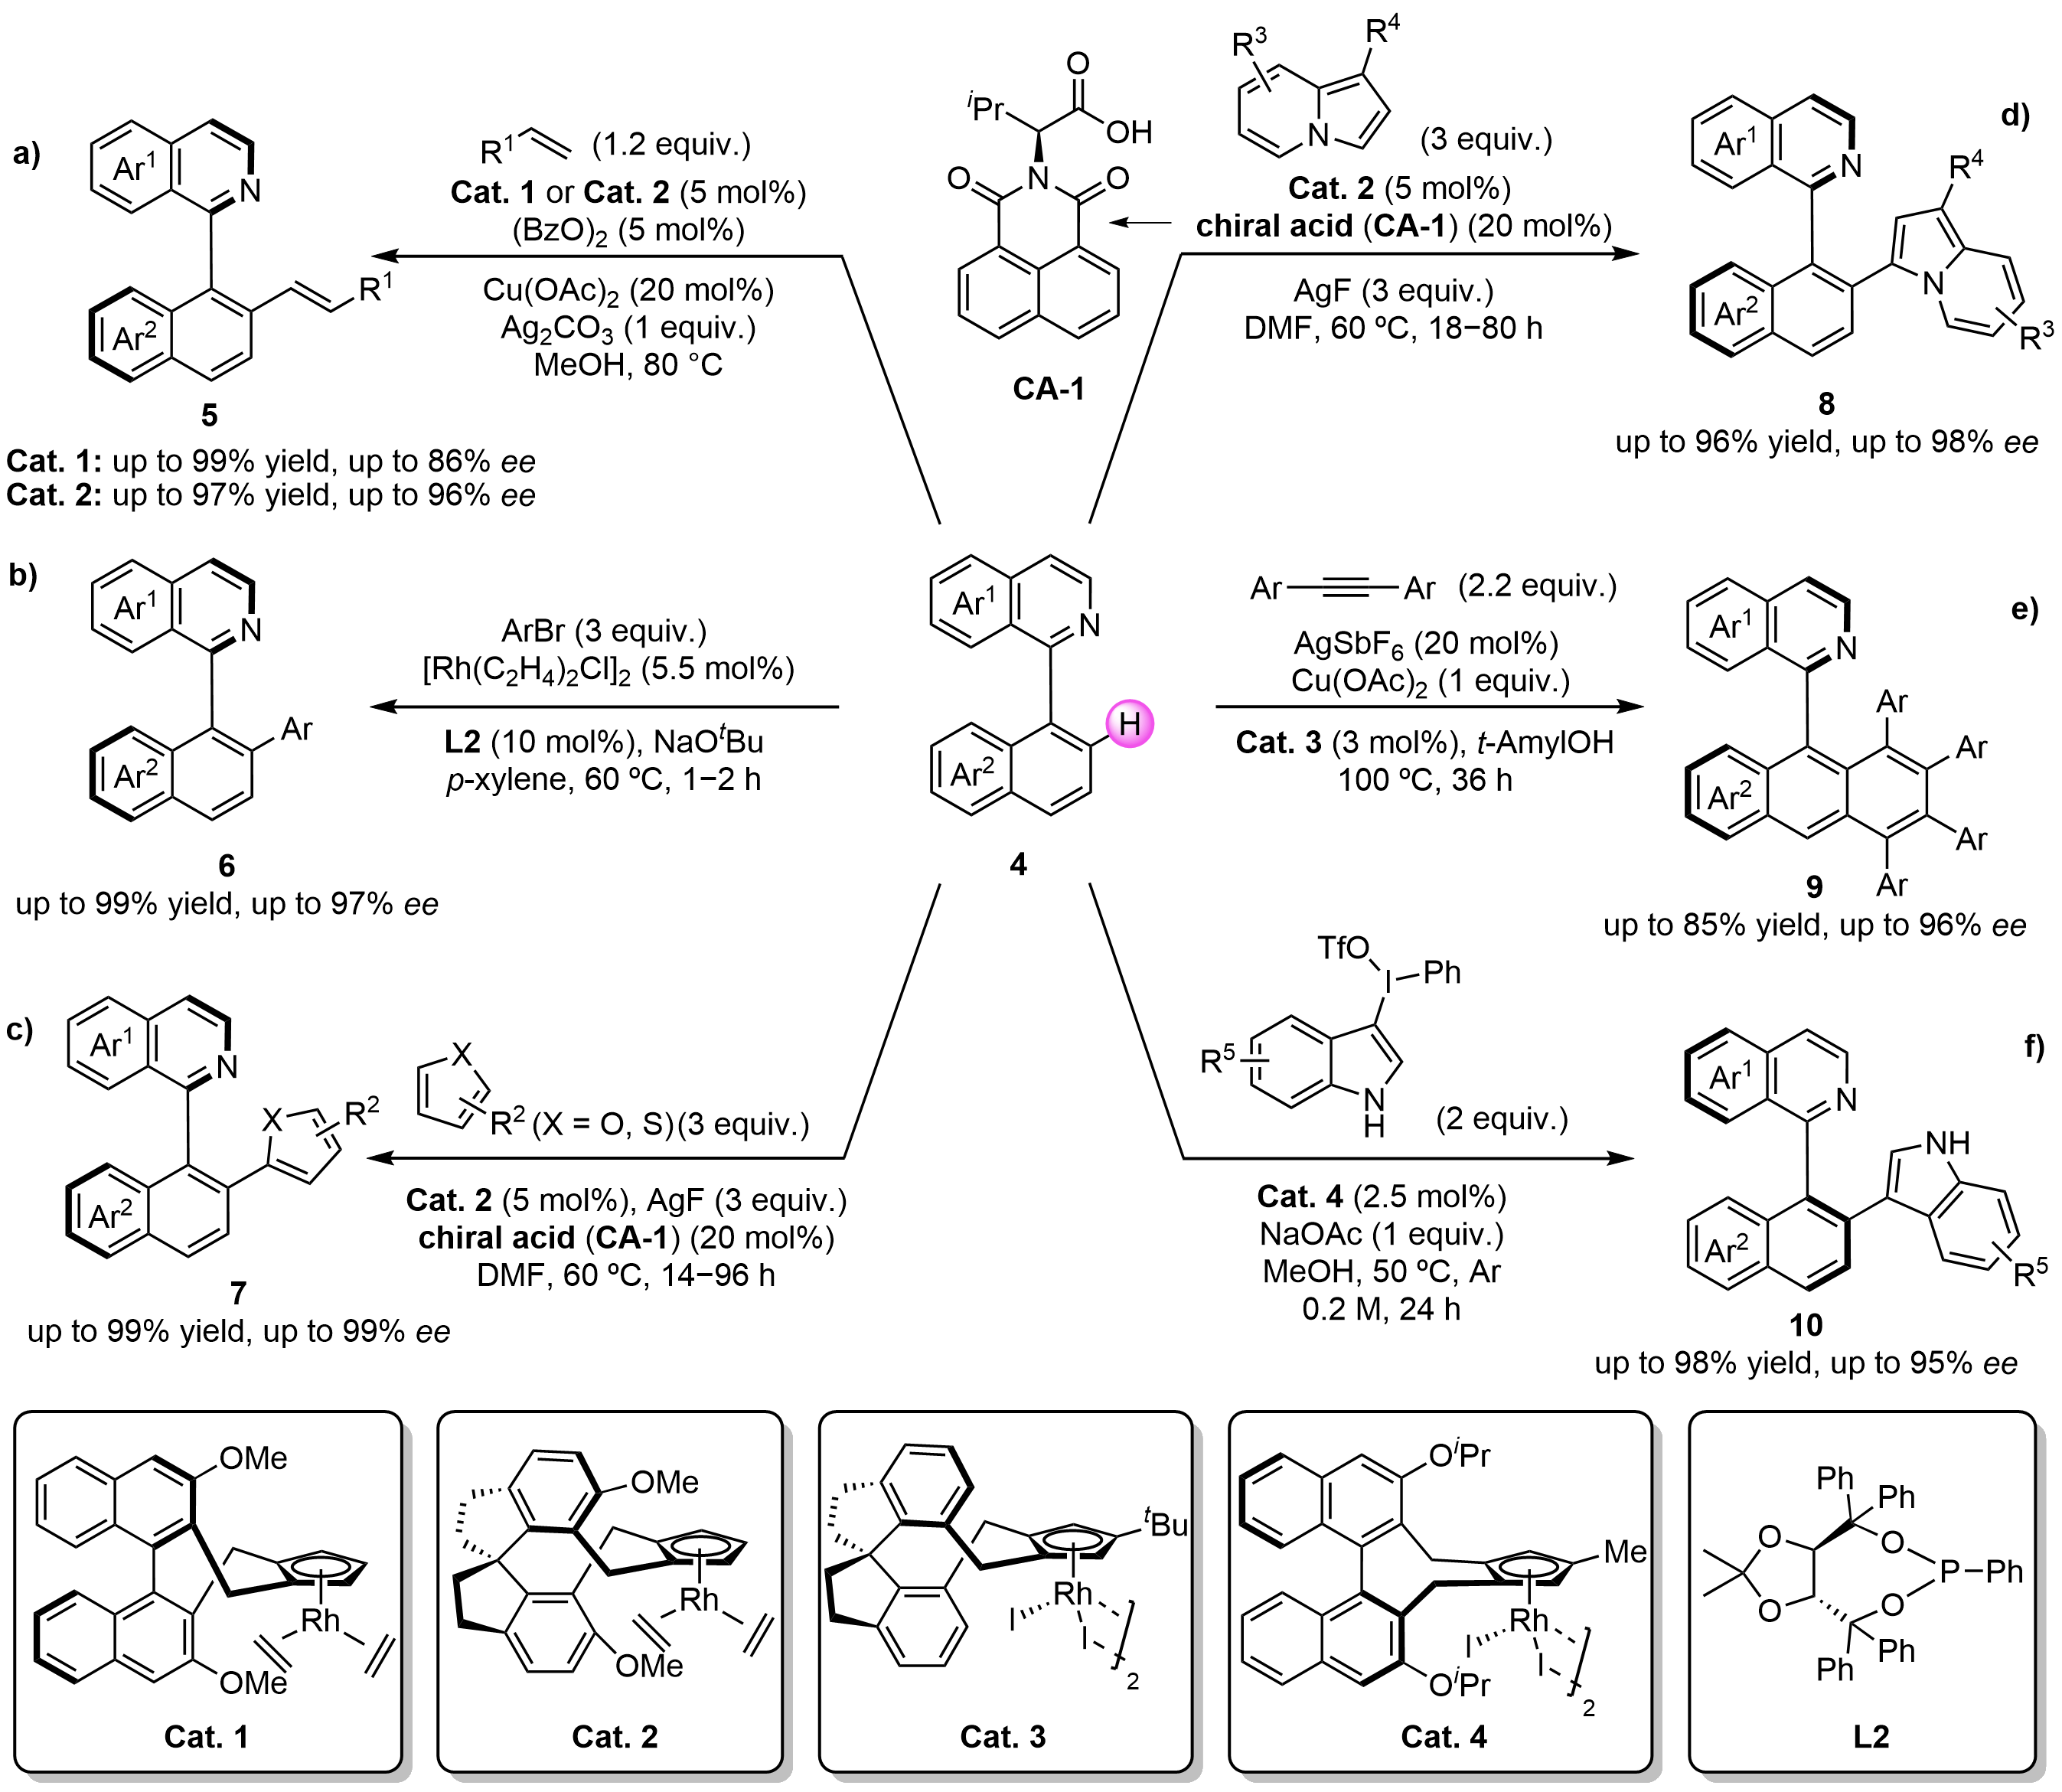
In 2014 and 2016, You and co-workers utilized pyridine-directed C−H activation strategy to develop the rhodium-catalyzed asymmetric cross-dehydrogenative coupling (CDC)[26-28] of biaryl compounds with alkenes (Scheme 6a)[29]. By employing an anionic chiral ligand of cyclopentadienyl (Cp) in a rhodium complex (Cat. 1), axially chiral products 5 were delivered in excellent yields with good enantioselectivity, presenting a broad range of substrates with good functional-group (FG) tolerance. Moreover, a higher enantioselectivity of the same product was observed under the catalysis of rhodium complex (Cat. 2) based on a chiral spirocyclic Cp-ligand[30]. The presence of the spiro skeleton in Cat. 2 facilitates a more effective extension of the methoxy group towards the rhodium center, potentially contributing to an enhanced chirality transfer implemented by the catalyst. In 2019, the You group used a rhodium catalyst, composed of [Rh(C2H4)2Cl]2 and TADDOL-derived monodentate phosphine ligands to achieve the pyridine-directed C−H arylation reaction with bromoarenes, affording axially chiral 1-arylisoquinolines in high yield with excellent enantioselectivity (Scheme 6b)[31].
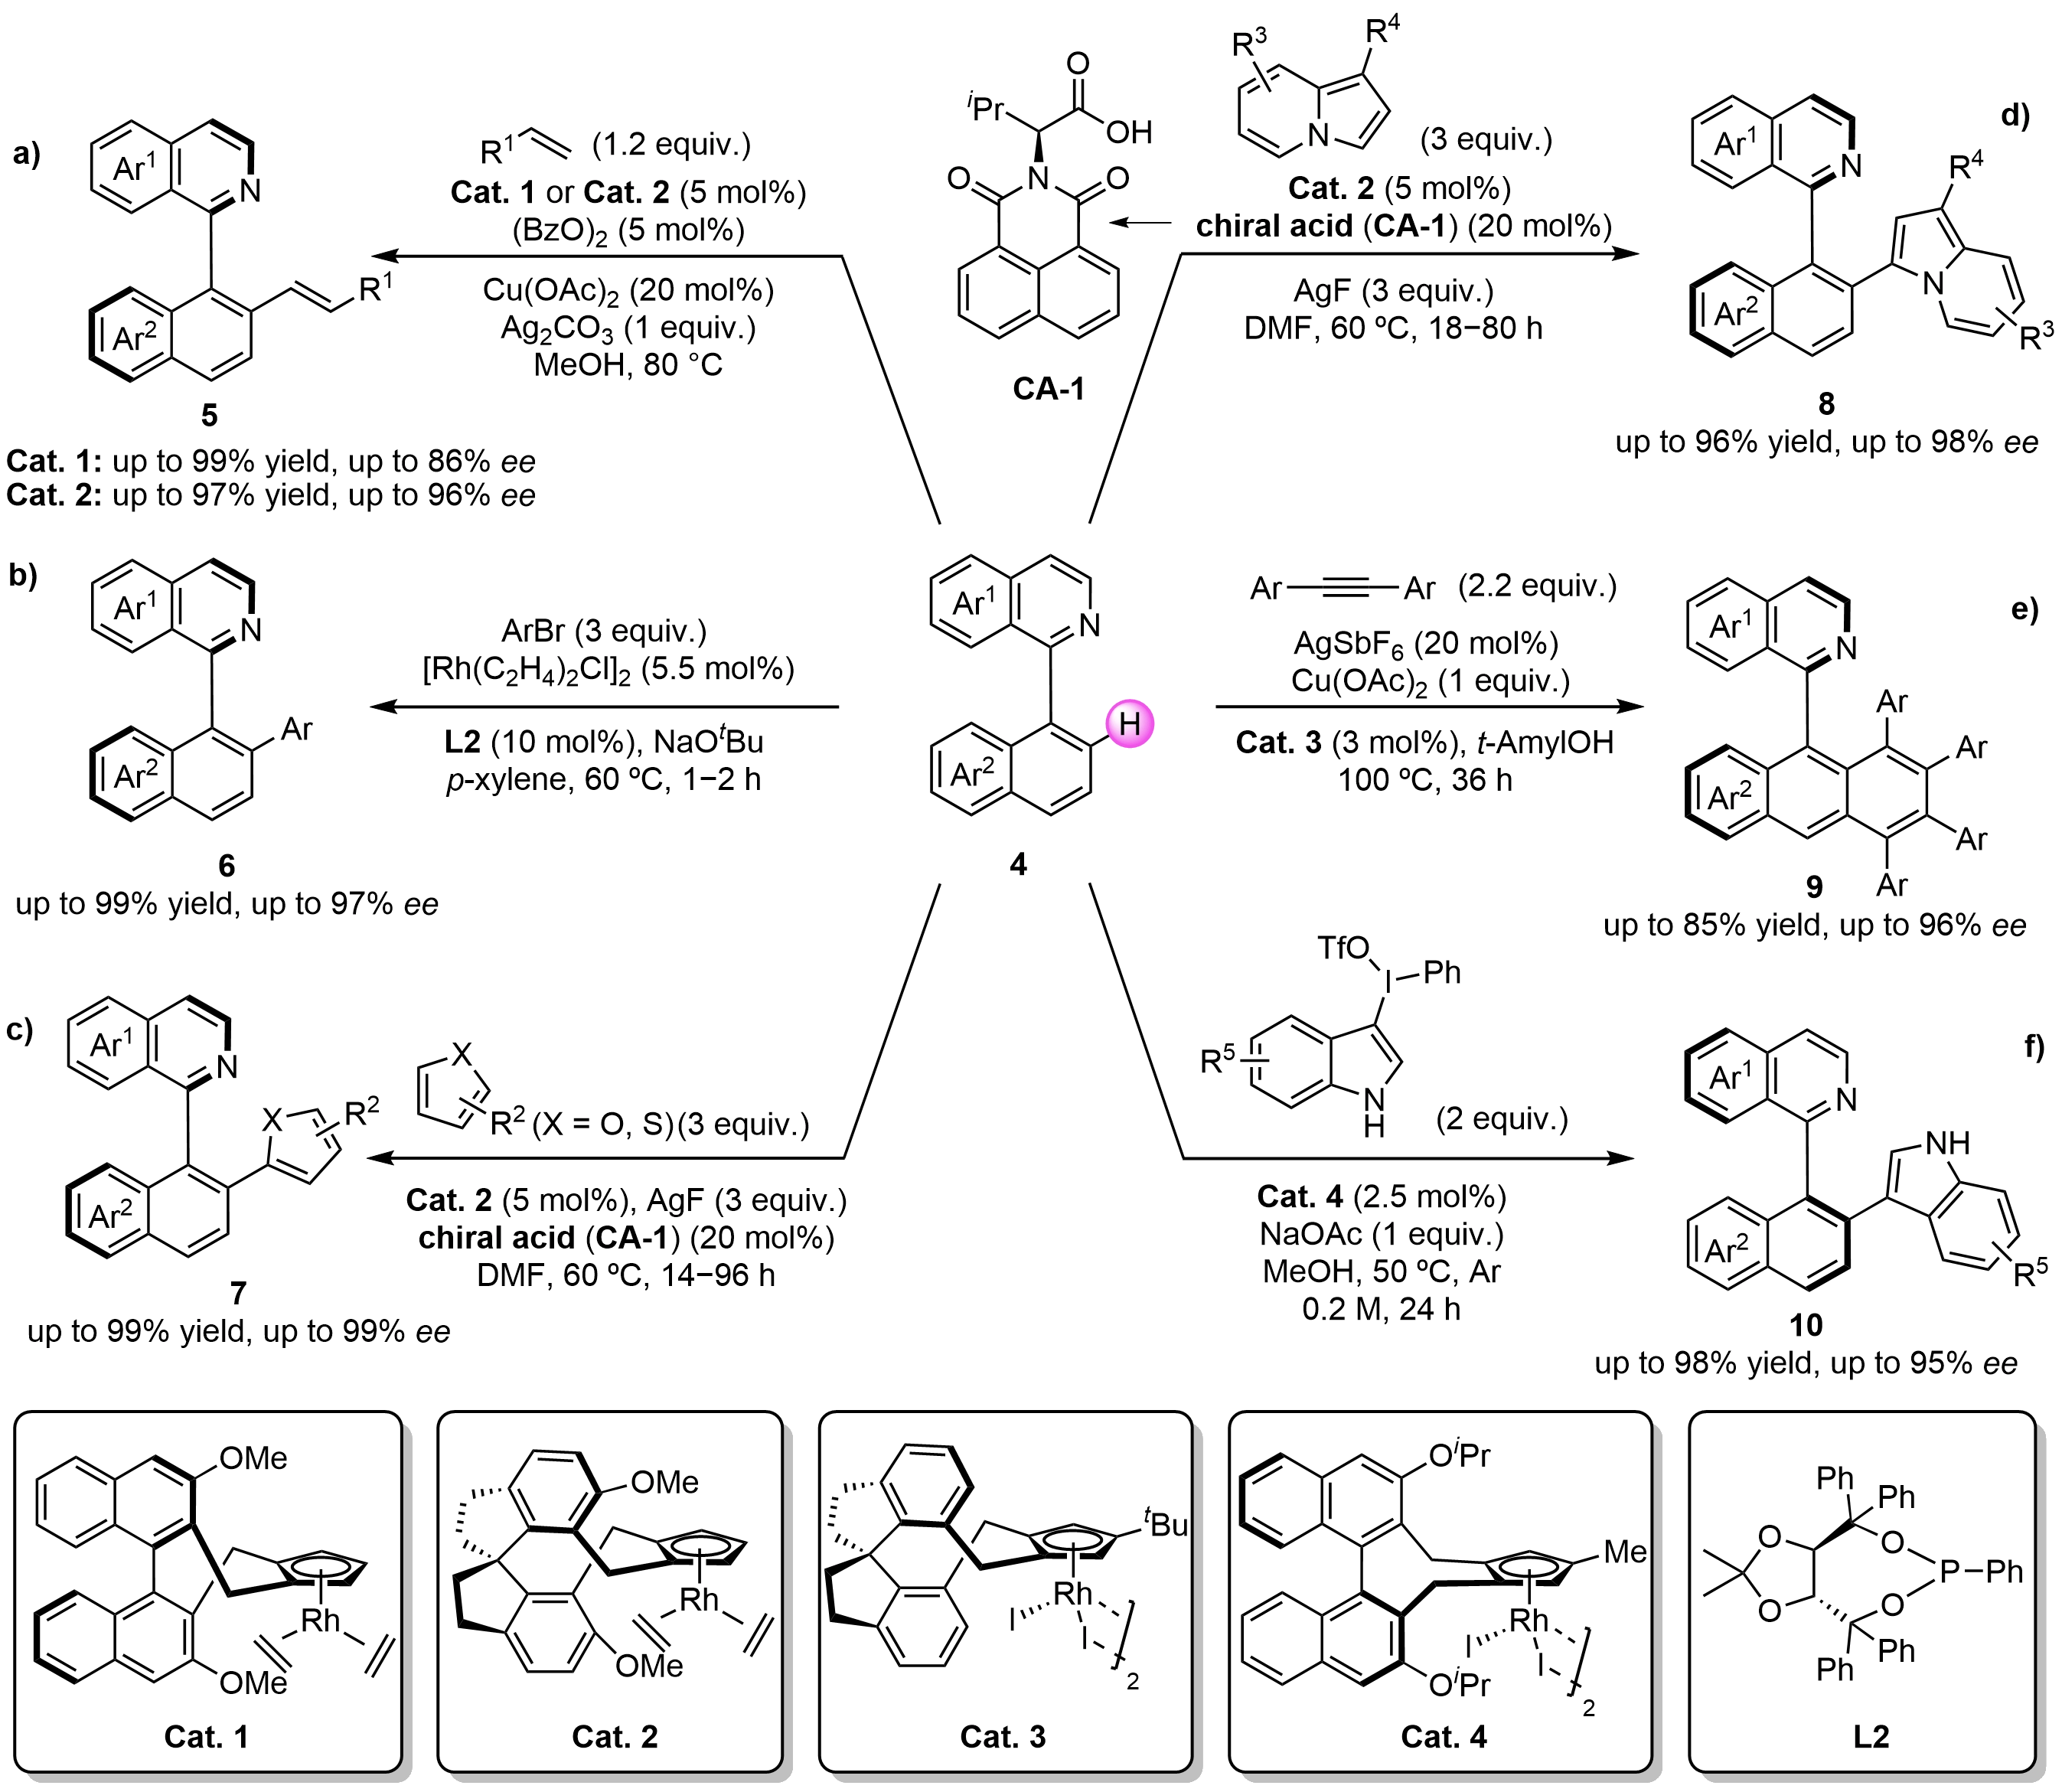
Scheme 6. Construction of C−C axial chirality by rhodium/iridium-catalyzed C−H functionalization of 1-arylisoquinoline.
In 2020, the same group effectively applied the previously developed chiral rhodium catalyst (Cat. 2) in cooperation with a chiral carboxylic acid (CA-1) to accomplish the C−H cross-coupling reaction of 1-arylisoquinoline derivatives with electron-rich heteroarenes (Scheme 6c)[32]. Subsequently, building upon the aforementioned research, they exploited indolizines as the nucleophile with a similar DKR strategy (Scheme 6d)[33]. Under these conditions, stoichiometric amounts of oxidant (AgF) were indispensable.
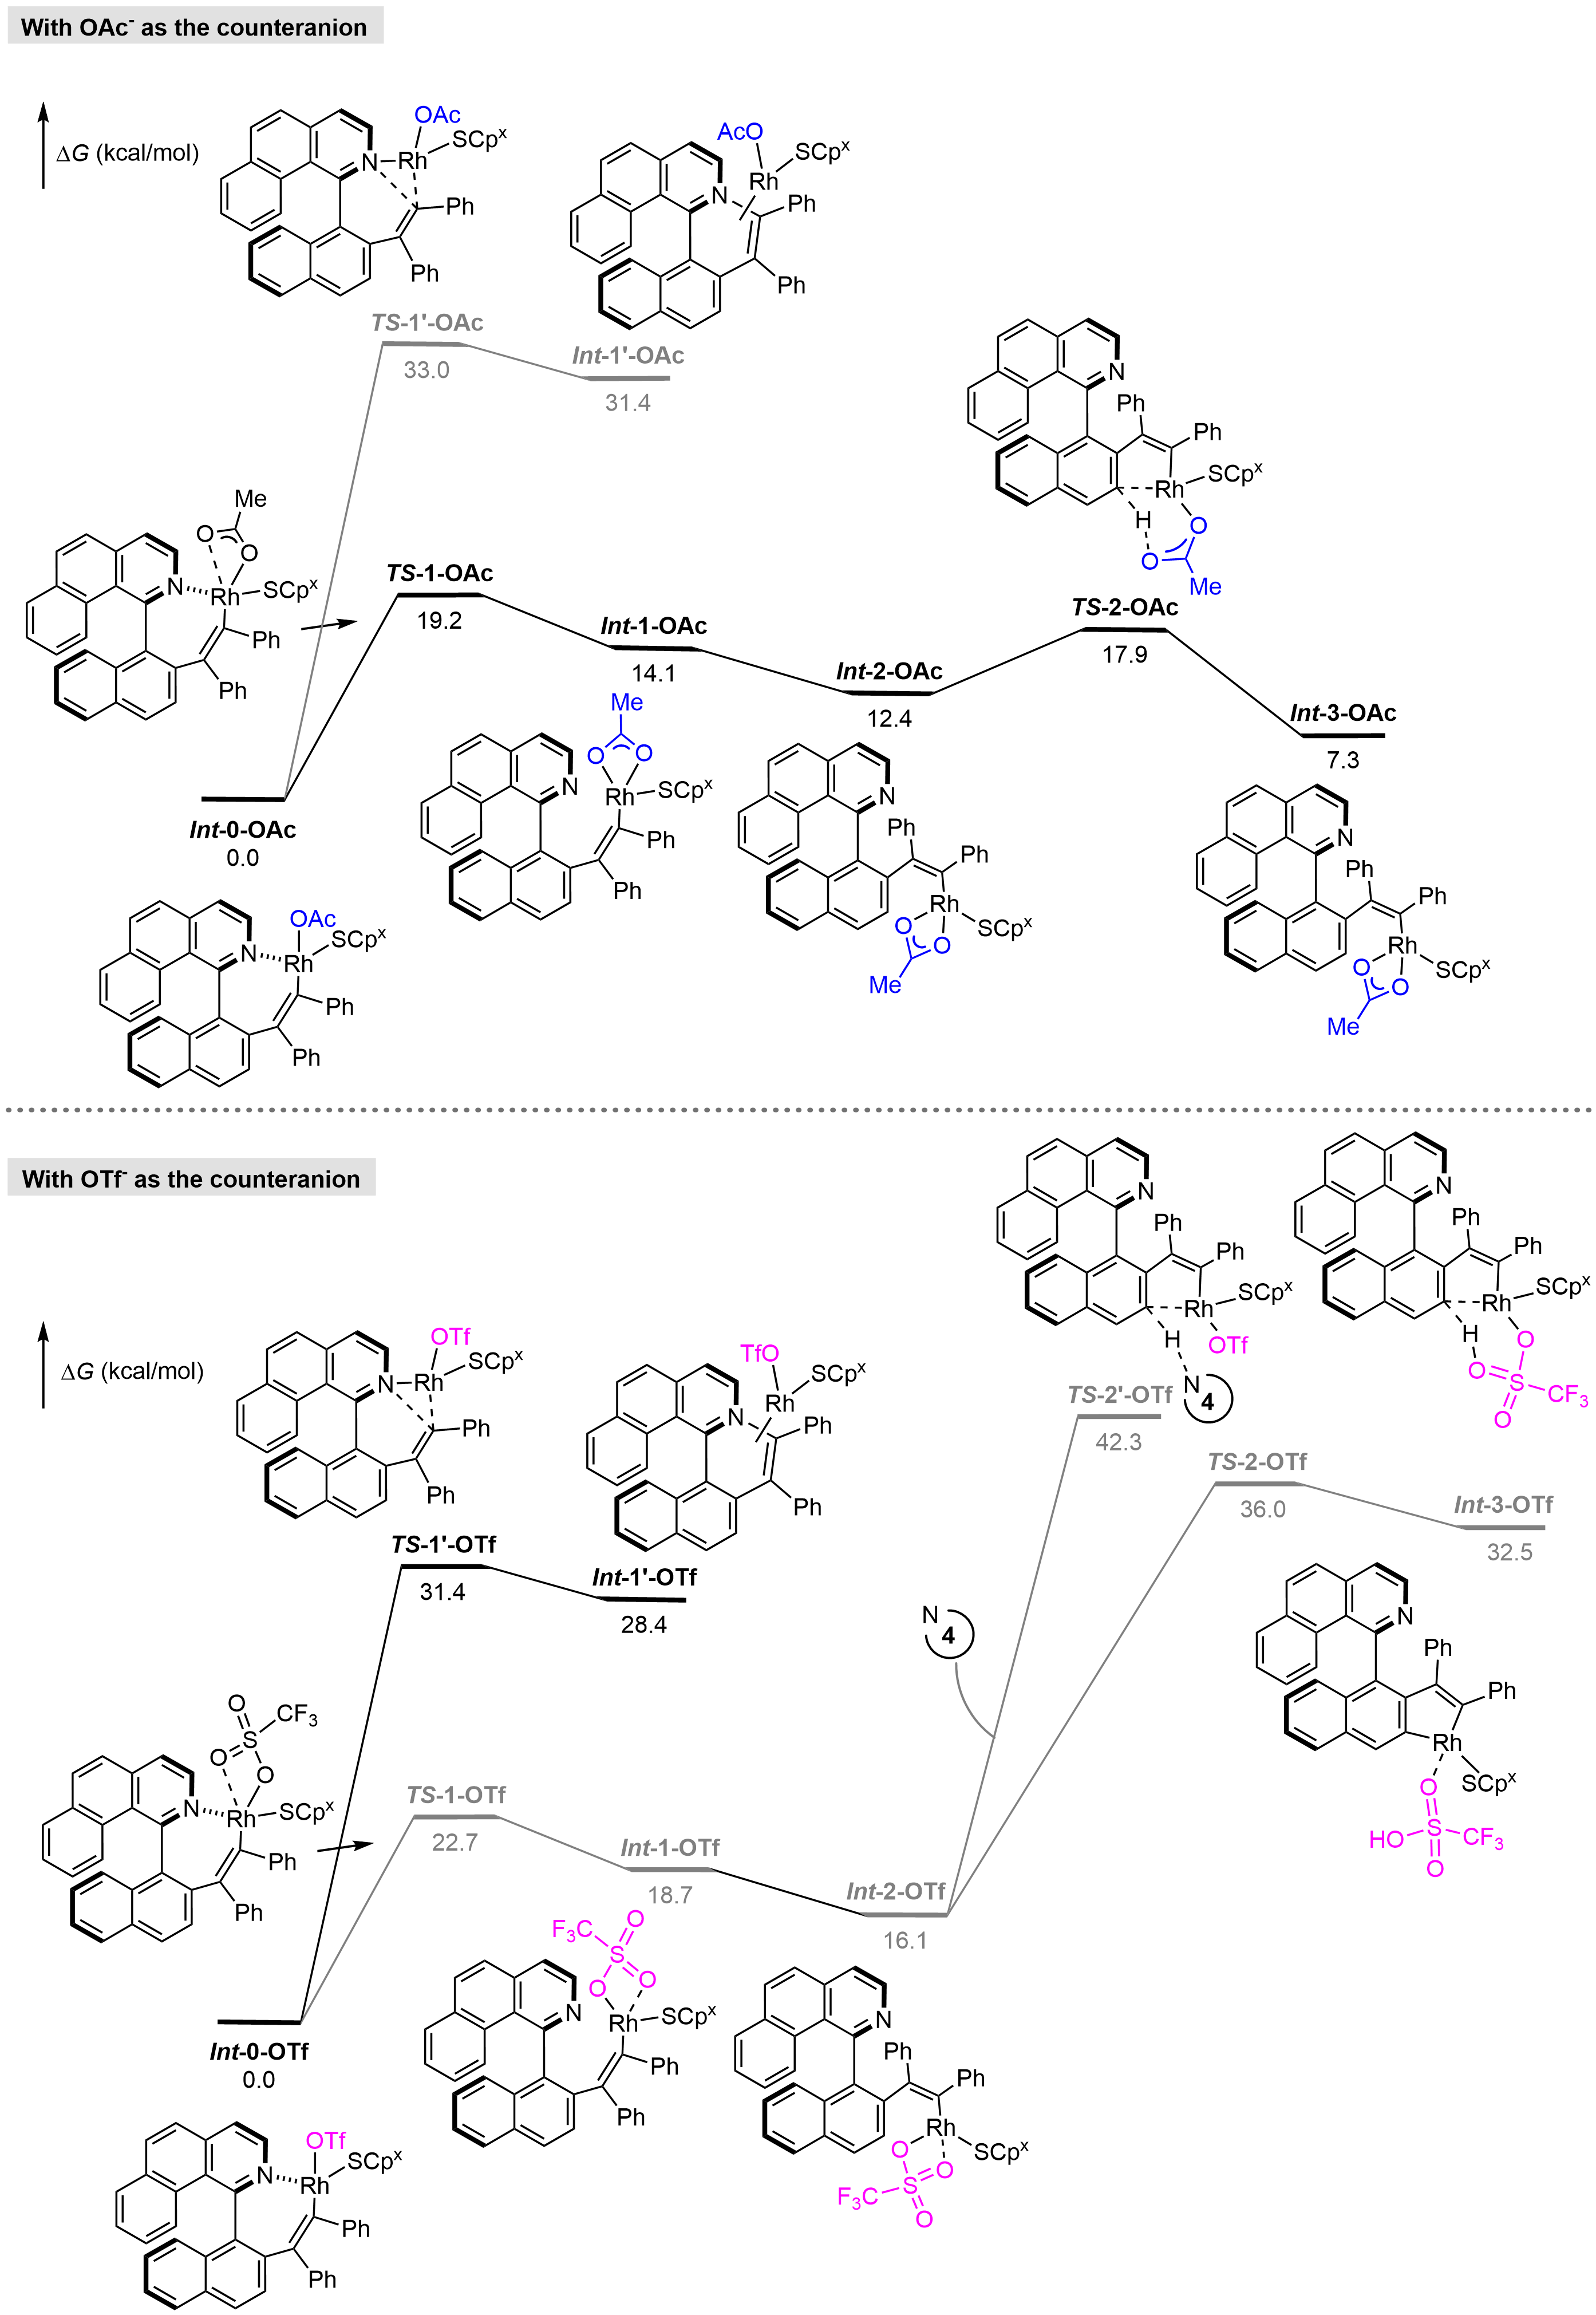
In 2023, You and co-workers further disclosed a new rhodium-catalyzed asymmetric Satoh-Miura reaction to access axially chiral compounds 9 bearing an extensive aromatic system (Scheme 6e)[34]. Control experiments demonstrated a significant influence of the counteranion of the metal on the reaction outcomes. The utilization of an anion with high alkalinity and coordination ability preferentially proceeds via the Satoh-Miura reaction pathway. In contrast, the employment of a weakly basic and non-coordinating anion favors the formation of azoniahelicene[35] products. To demonstrate the mechanism of the reaction pathway guided by the dominant counteranion of the system, they also conducted density functional theory (DFT) calculations on the reactions when OAc- or OTf- were the main dominant counteranion in the system. With OAc- as the counterion, the Satoh-Miura reaction pathway is more energetically favorable compared to the formation of the azoniahelicene product. In the reaction pathway for the Satoh-Miura reaction, their energies were much lower than that of the C−N reductive elimination transition state (TS-1'-OAc, 33.0 kcal/mol). In contrast, when using OTf- as the counteranion, the C−N reductive elimination preferentially occurred via the transition state (TS-1'-OTf, 31.4 kcal/mol) to form the azoniahelicene product, given that the second C−H activation in the Satoh-Miura reaction pathway is rather difficult (Scheme 7).
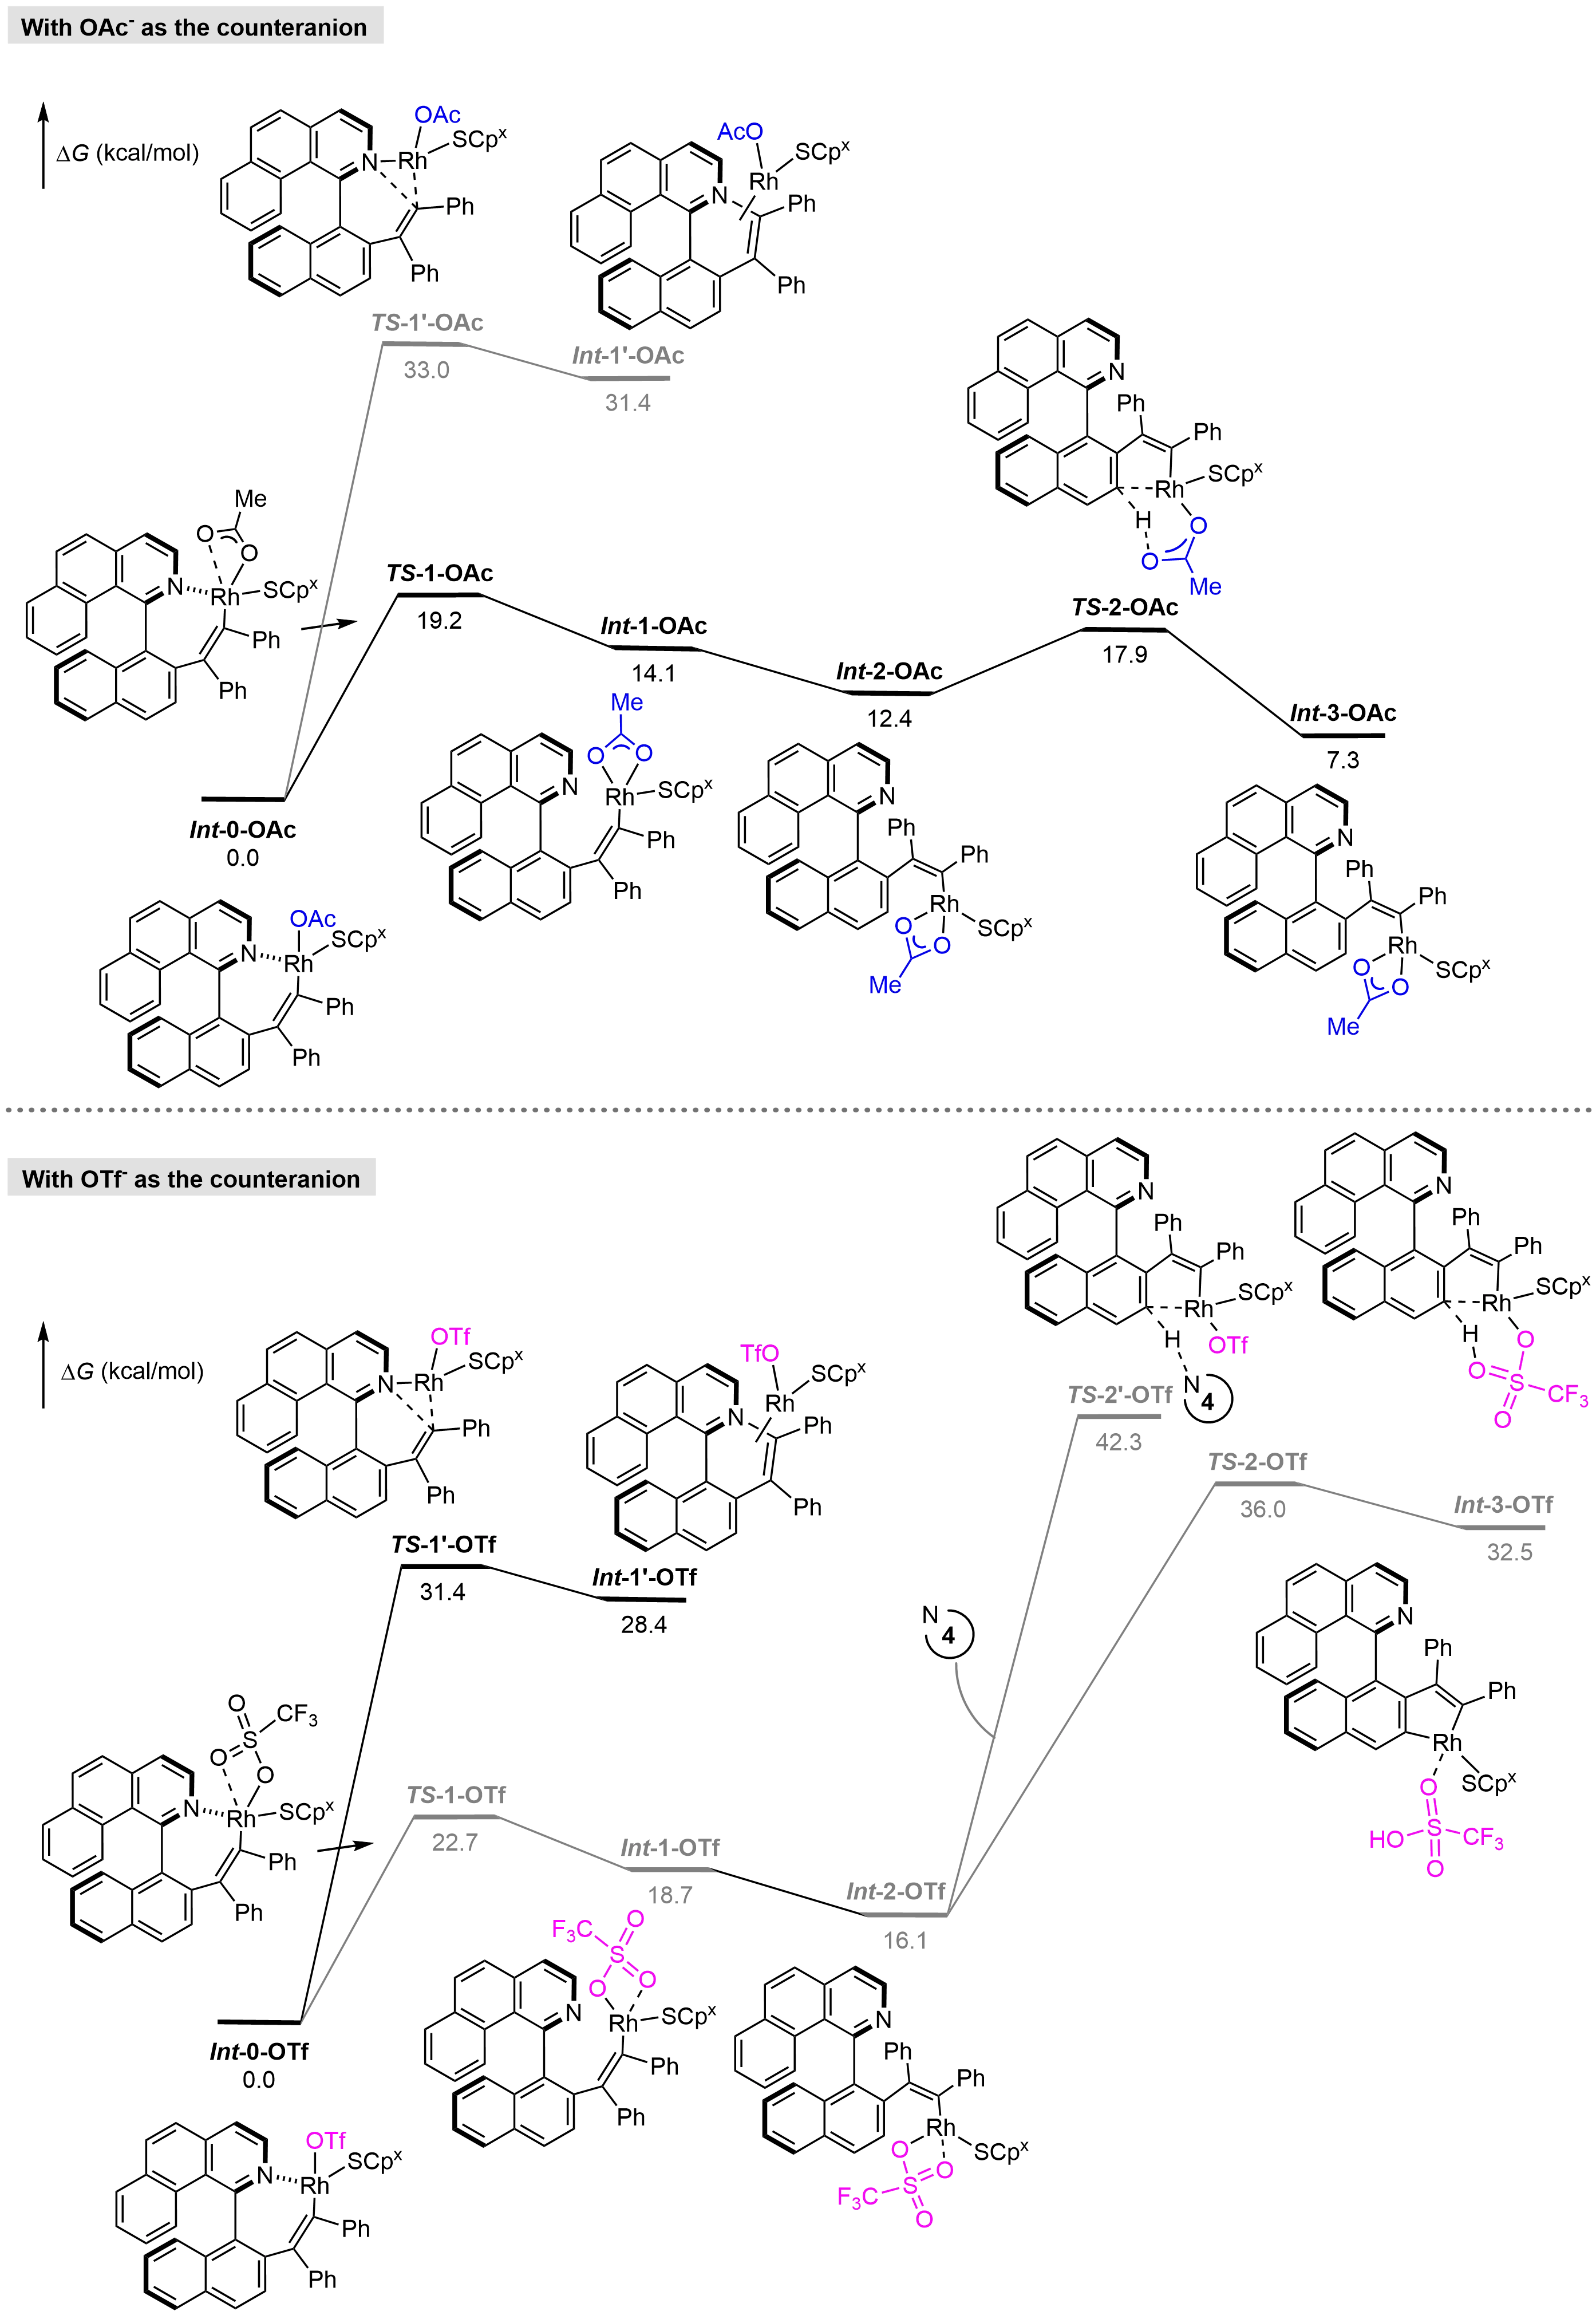
Scheme 7. The energy profile of the reaction involving OAc- and OTf- as the counteranion by DFT calculations. DFT: density functional theory.
Additionally, the authors applied internal alkynes to achieve the alkenylation of 1-arylisoquinoline under iridium catalysis[36]. In 2023 and 2024, they further accomplished the rhodium-catalyzed asymmetric C−H indolylation reaction with hypervalent iodine reagents, enabling the facile connection of an axially chiral 1-arylisoquinoline unit with an indole moiety under mild conditions. It is noteworthy that this DKR via asymmetric C−H indolylation presents exceptional efficiency (up to 98% yield) and enantioselectivity (up to 95% ee) (Scheme 6f)[37]. Similarly, the rhodium-catalyzed asymmetric C−H alkynylation, iodination, selenylation, and cyanation of 1-arylisoquinoline can also be realized with alkynyl-substituted hypervalent iodine reagents[38], N-iodosuccinimide (NIS)[39], organoselenide reagents[40], and cyanation reagents[41], respectively. In the aforementioned works, the authors calculated the rotational barriers of the representative products, thereby confirming their stable axial chirality.
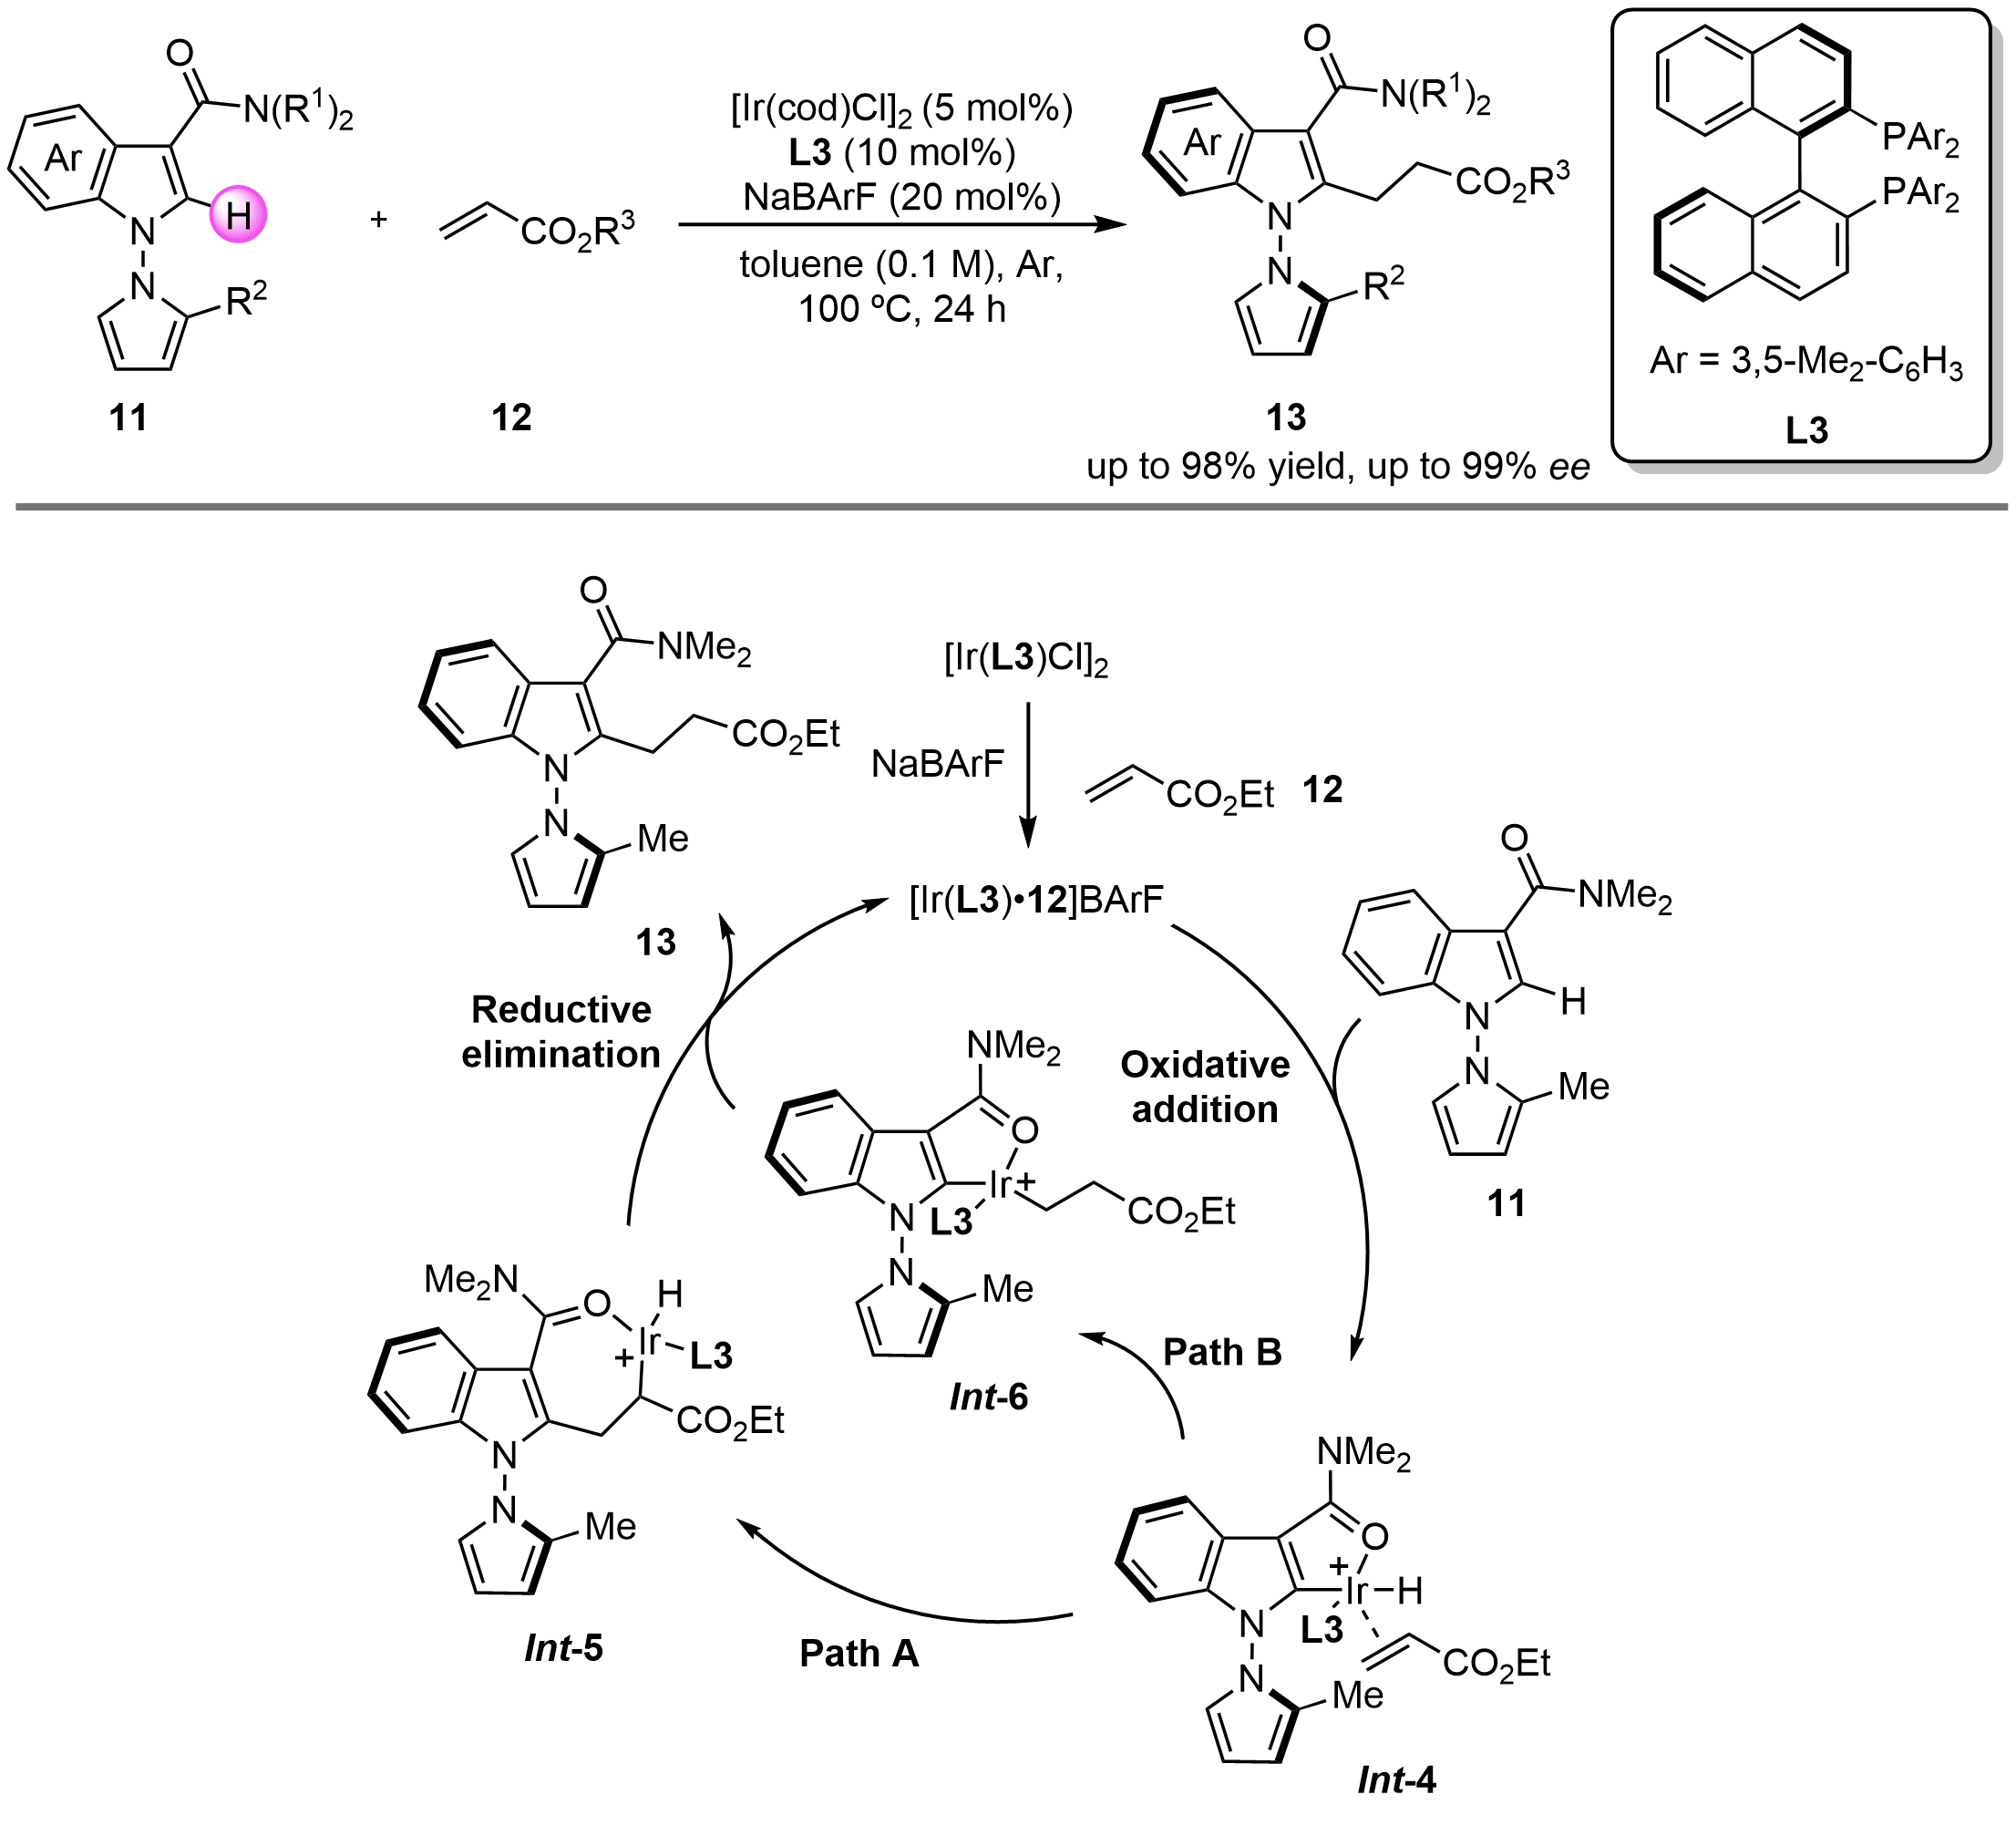
In addition to the establishment of C−C axial chirality, in 2023, You and co-workers elegantly achieved the construction of N−N axial chirality via amide-directed asymmetric C−H alkylation of indole-pyrrole adducts with acrylates enabled by iridium catalysis. The DKR process may undergo sequential oxidative addition, migratory insertion, and reductive elimination to complete the catalytic cycle, in which two possible paths were proposed for the step of migratory insertion, i.e., migration of the aryl group in Path A and hydride migration in Path B, respectively (Scheme 8)[42].
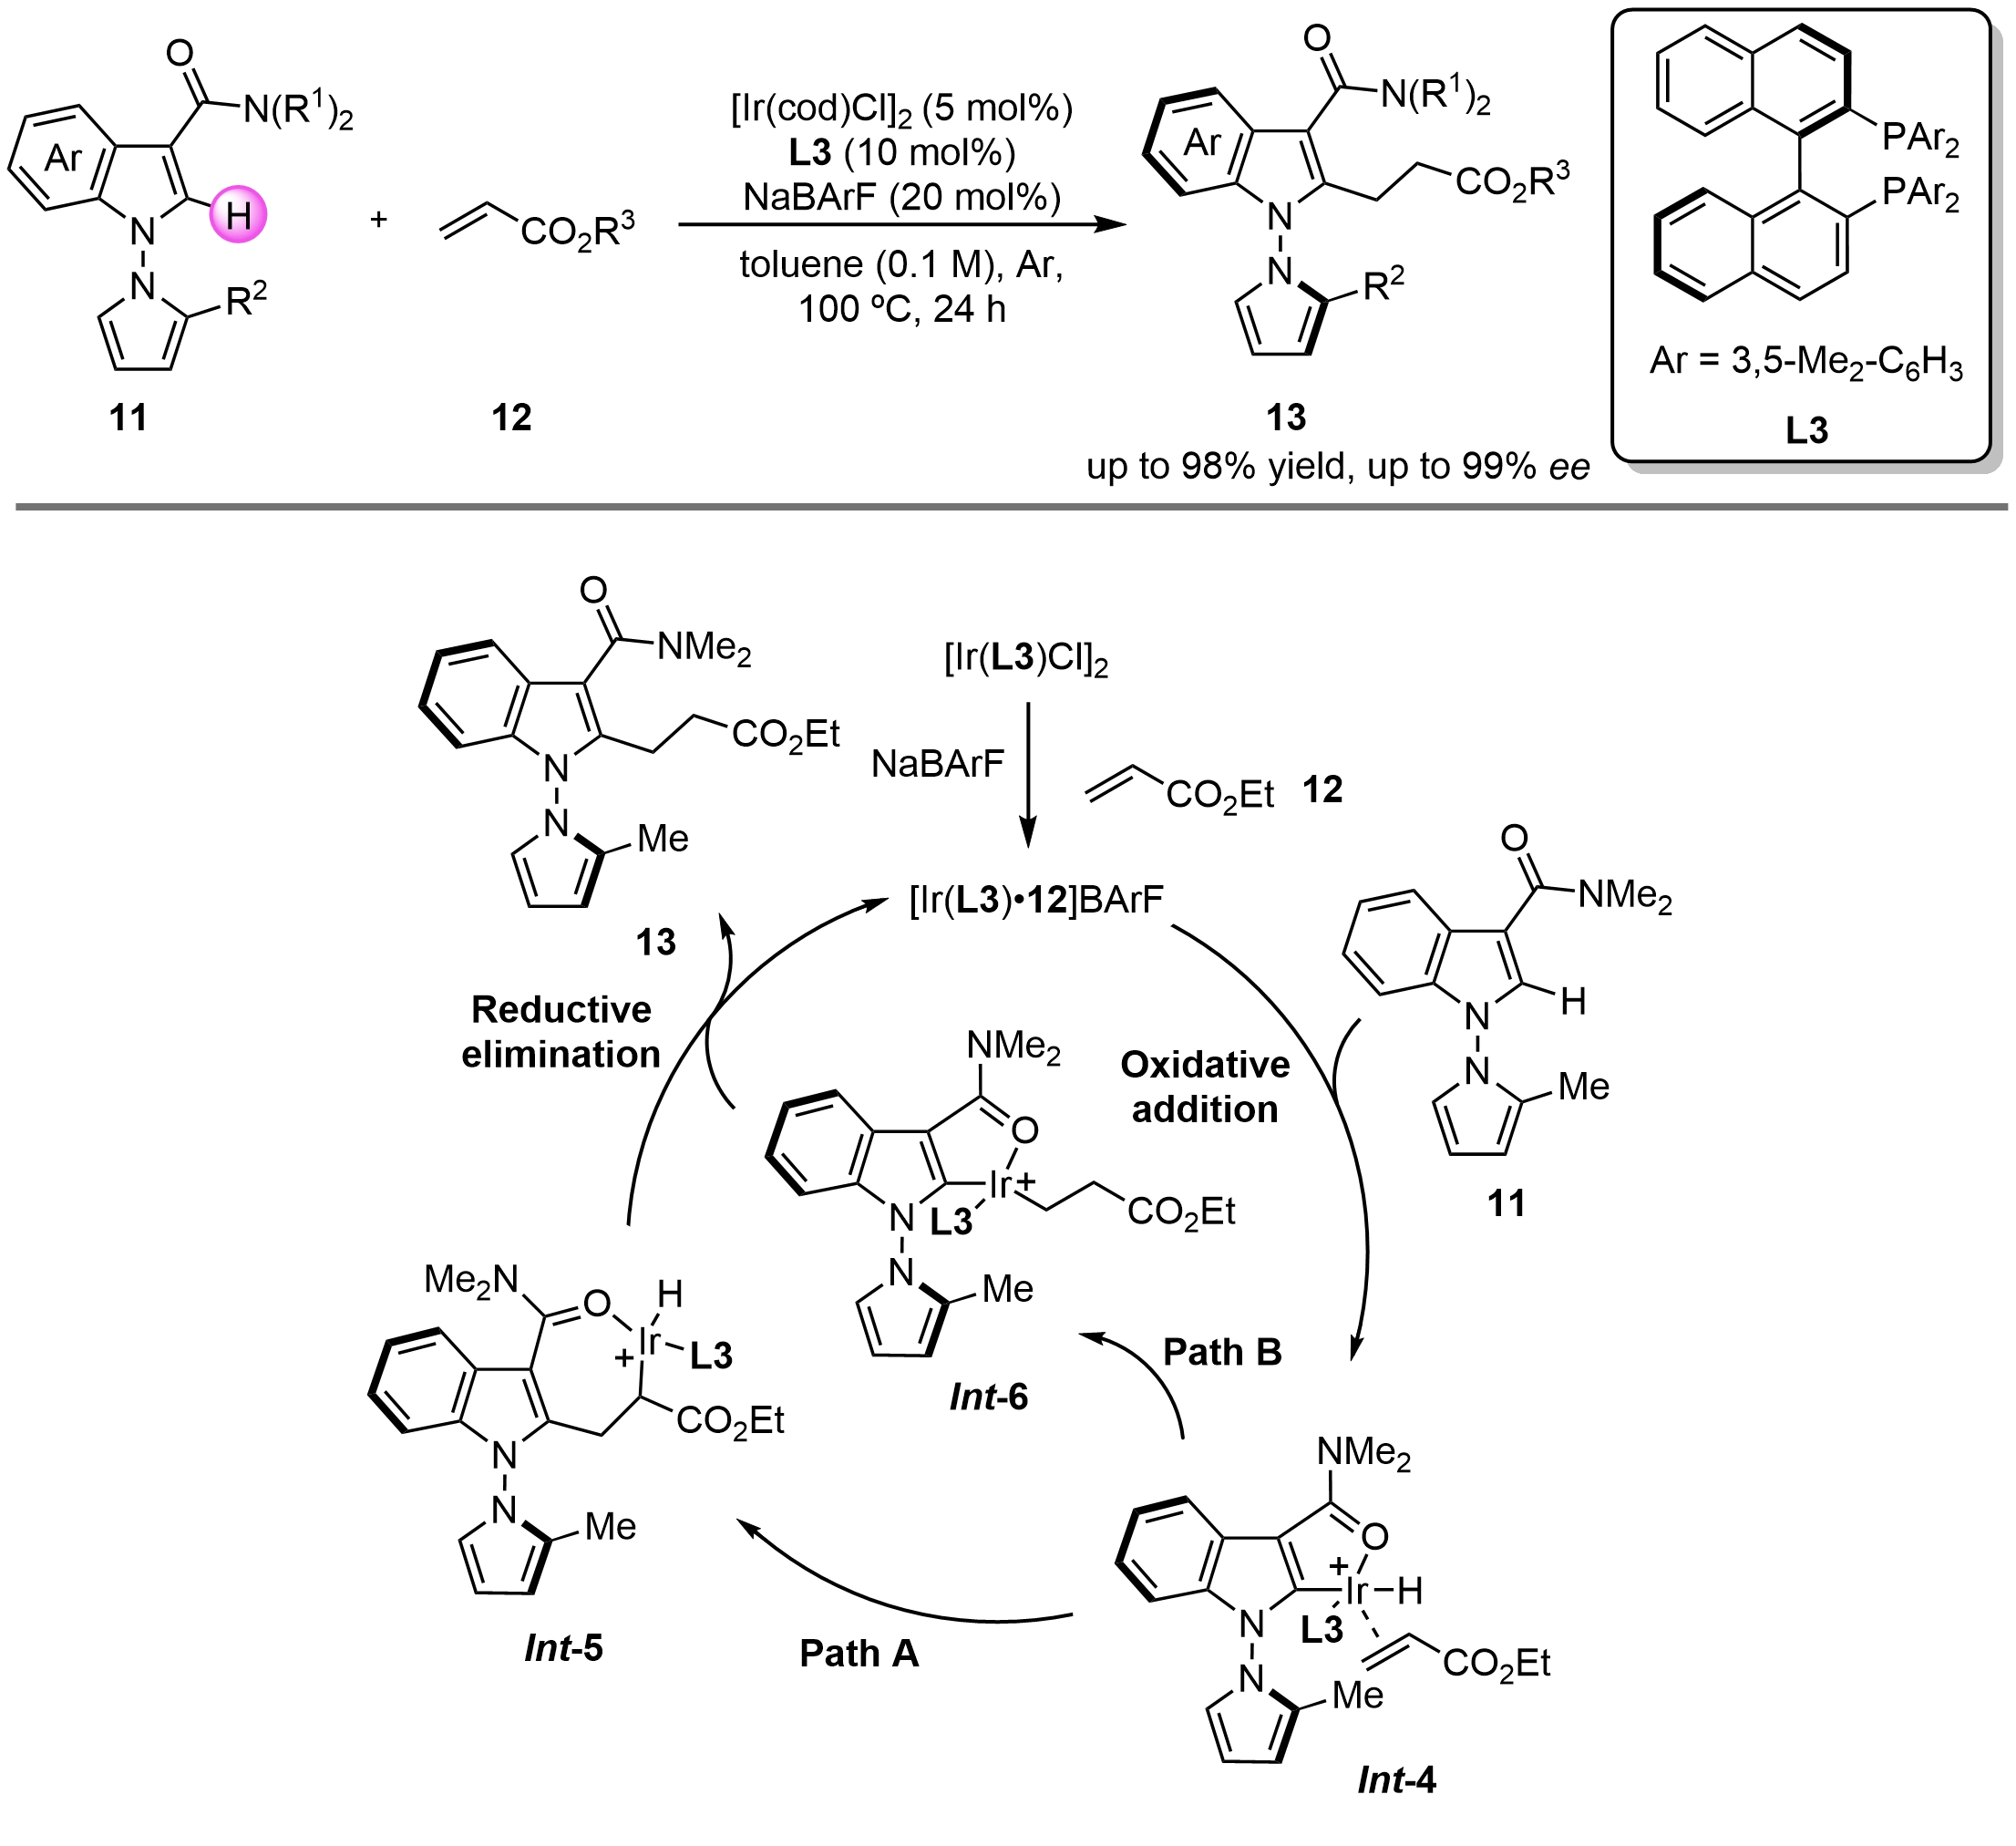
Scheme 8. Construction of N−N axial chirality via iridium-catalyzed asymmetric C−H alkylation of indole-pyrrole adducts with acrylates.
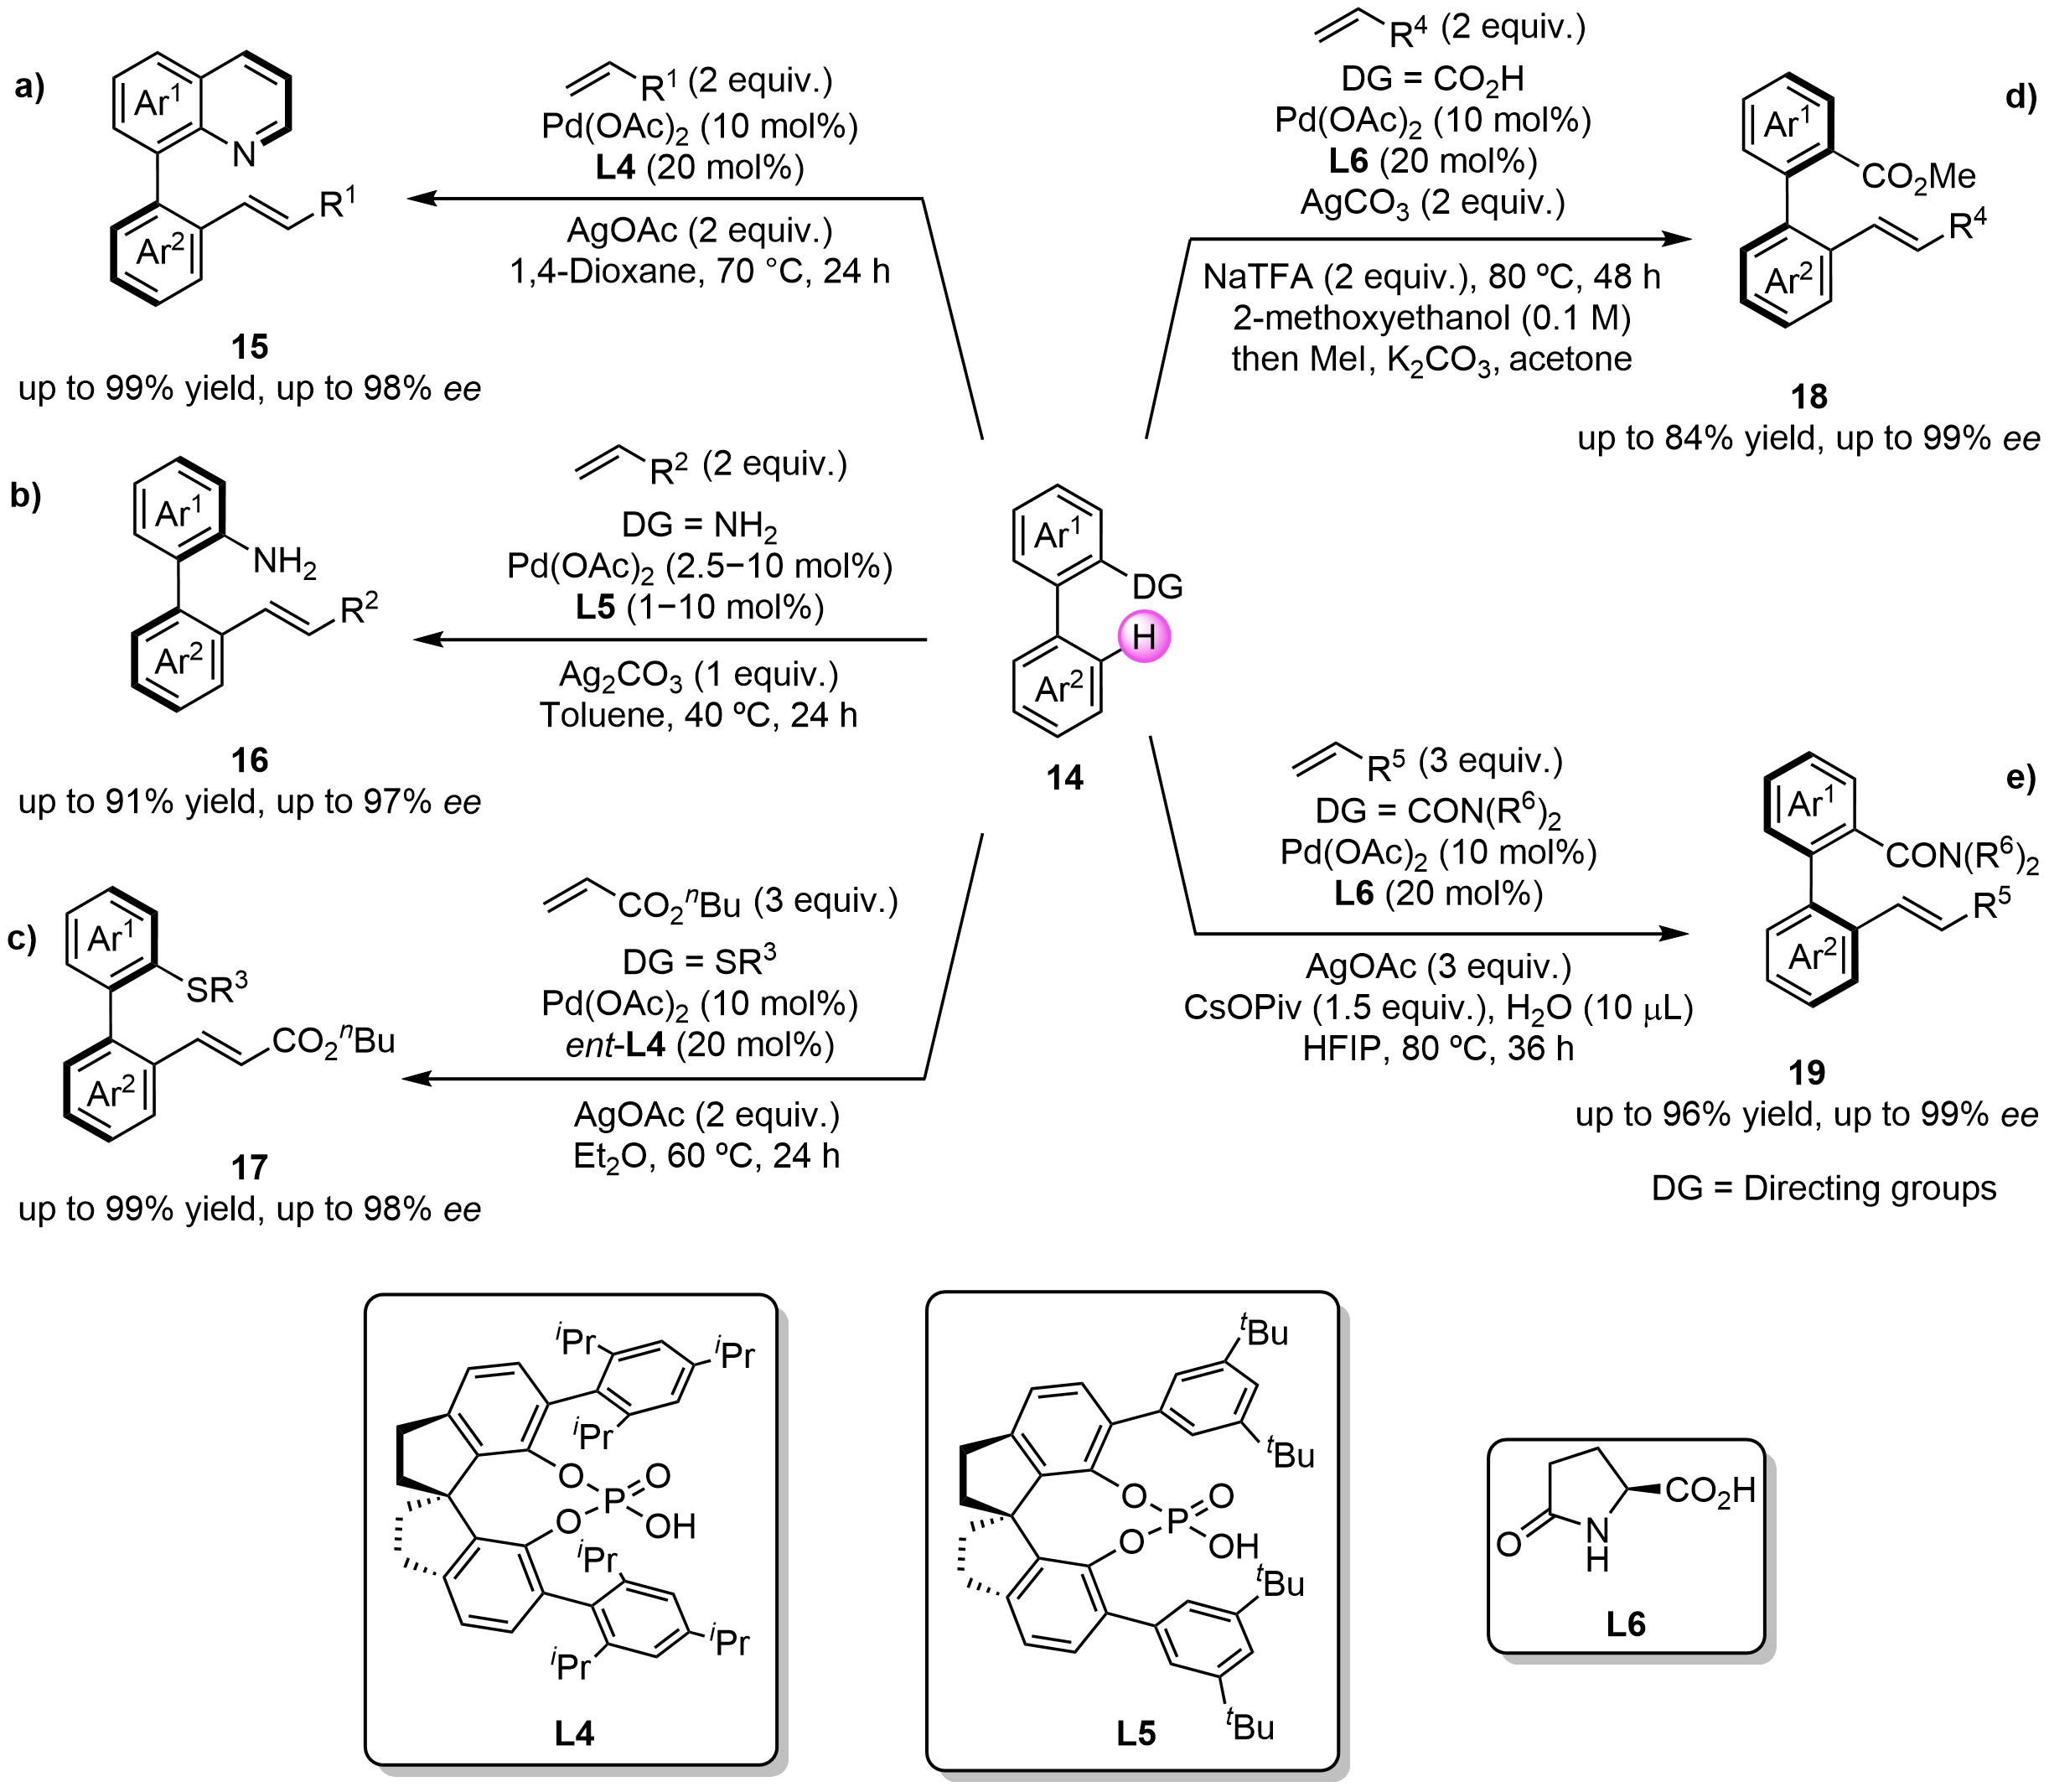
In 2019, the Shi group developed a palladium/chiral phosphoric acid-cocatalyzed asymmetric C−H Heck coupling approach. Quinoline as a directing group promoted the efficient oxidative olefination to present a class of axially chiral quinoline derivatives with high enantioselectivity (up to 98% ee, Scheme 9a)[43]. In this reaction, chiral phosphoric acid L4 combines with palladium acetate to form a chiral reaction cavity for chiral induction. Similarly, the iridium-catalyzed isoquinoline-directed C−H alkylation and silylation approaches were reported to construct axially chiral heterobiaryls by the Lassaletta[44], Ackermann[45], and He[46] groups independently. Subsequently, the Shi group utilized a more reactive primary amino group as the directing group to realize the asymmetric C−H olefination (Scheme 9b)[47,48] and allylation[49] reactions, leading to a series of olefinated and allylated axially chiral skeletons, respectively. The presence of a free amine unit facilitates significant potential for subsequent synthetic applications. From 2022 to 2024, the same group also demonstrated that thioether[50], carboxylic acid[51], and amide[52] groups are also efficient directing groups to trigger the Heck reaction to afford corresponding products in excellent yields with high enantioselectivity (Scheme 9c,d,e). The authors have also devised a cobalt-catalyzed C−H arylation reaction with arylboronic acids by using a bidentate directing group of pyridinamide[53]. In addition, the Yang group utilized phosphine oxide as the directing group to promote an efficient access to axially chiral biaryl phosphine oxide compounds under the catalytic system of palladium and L-valine variant[54].
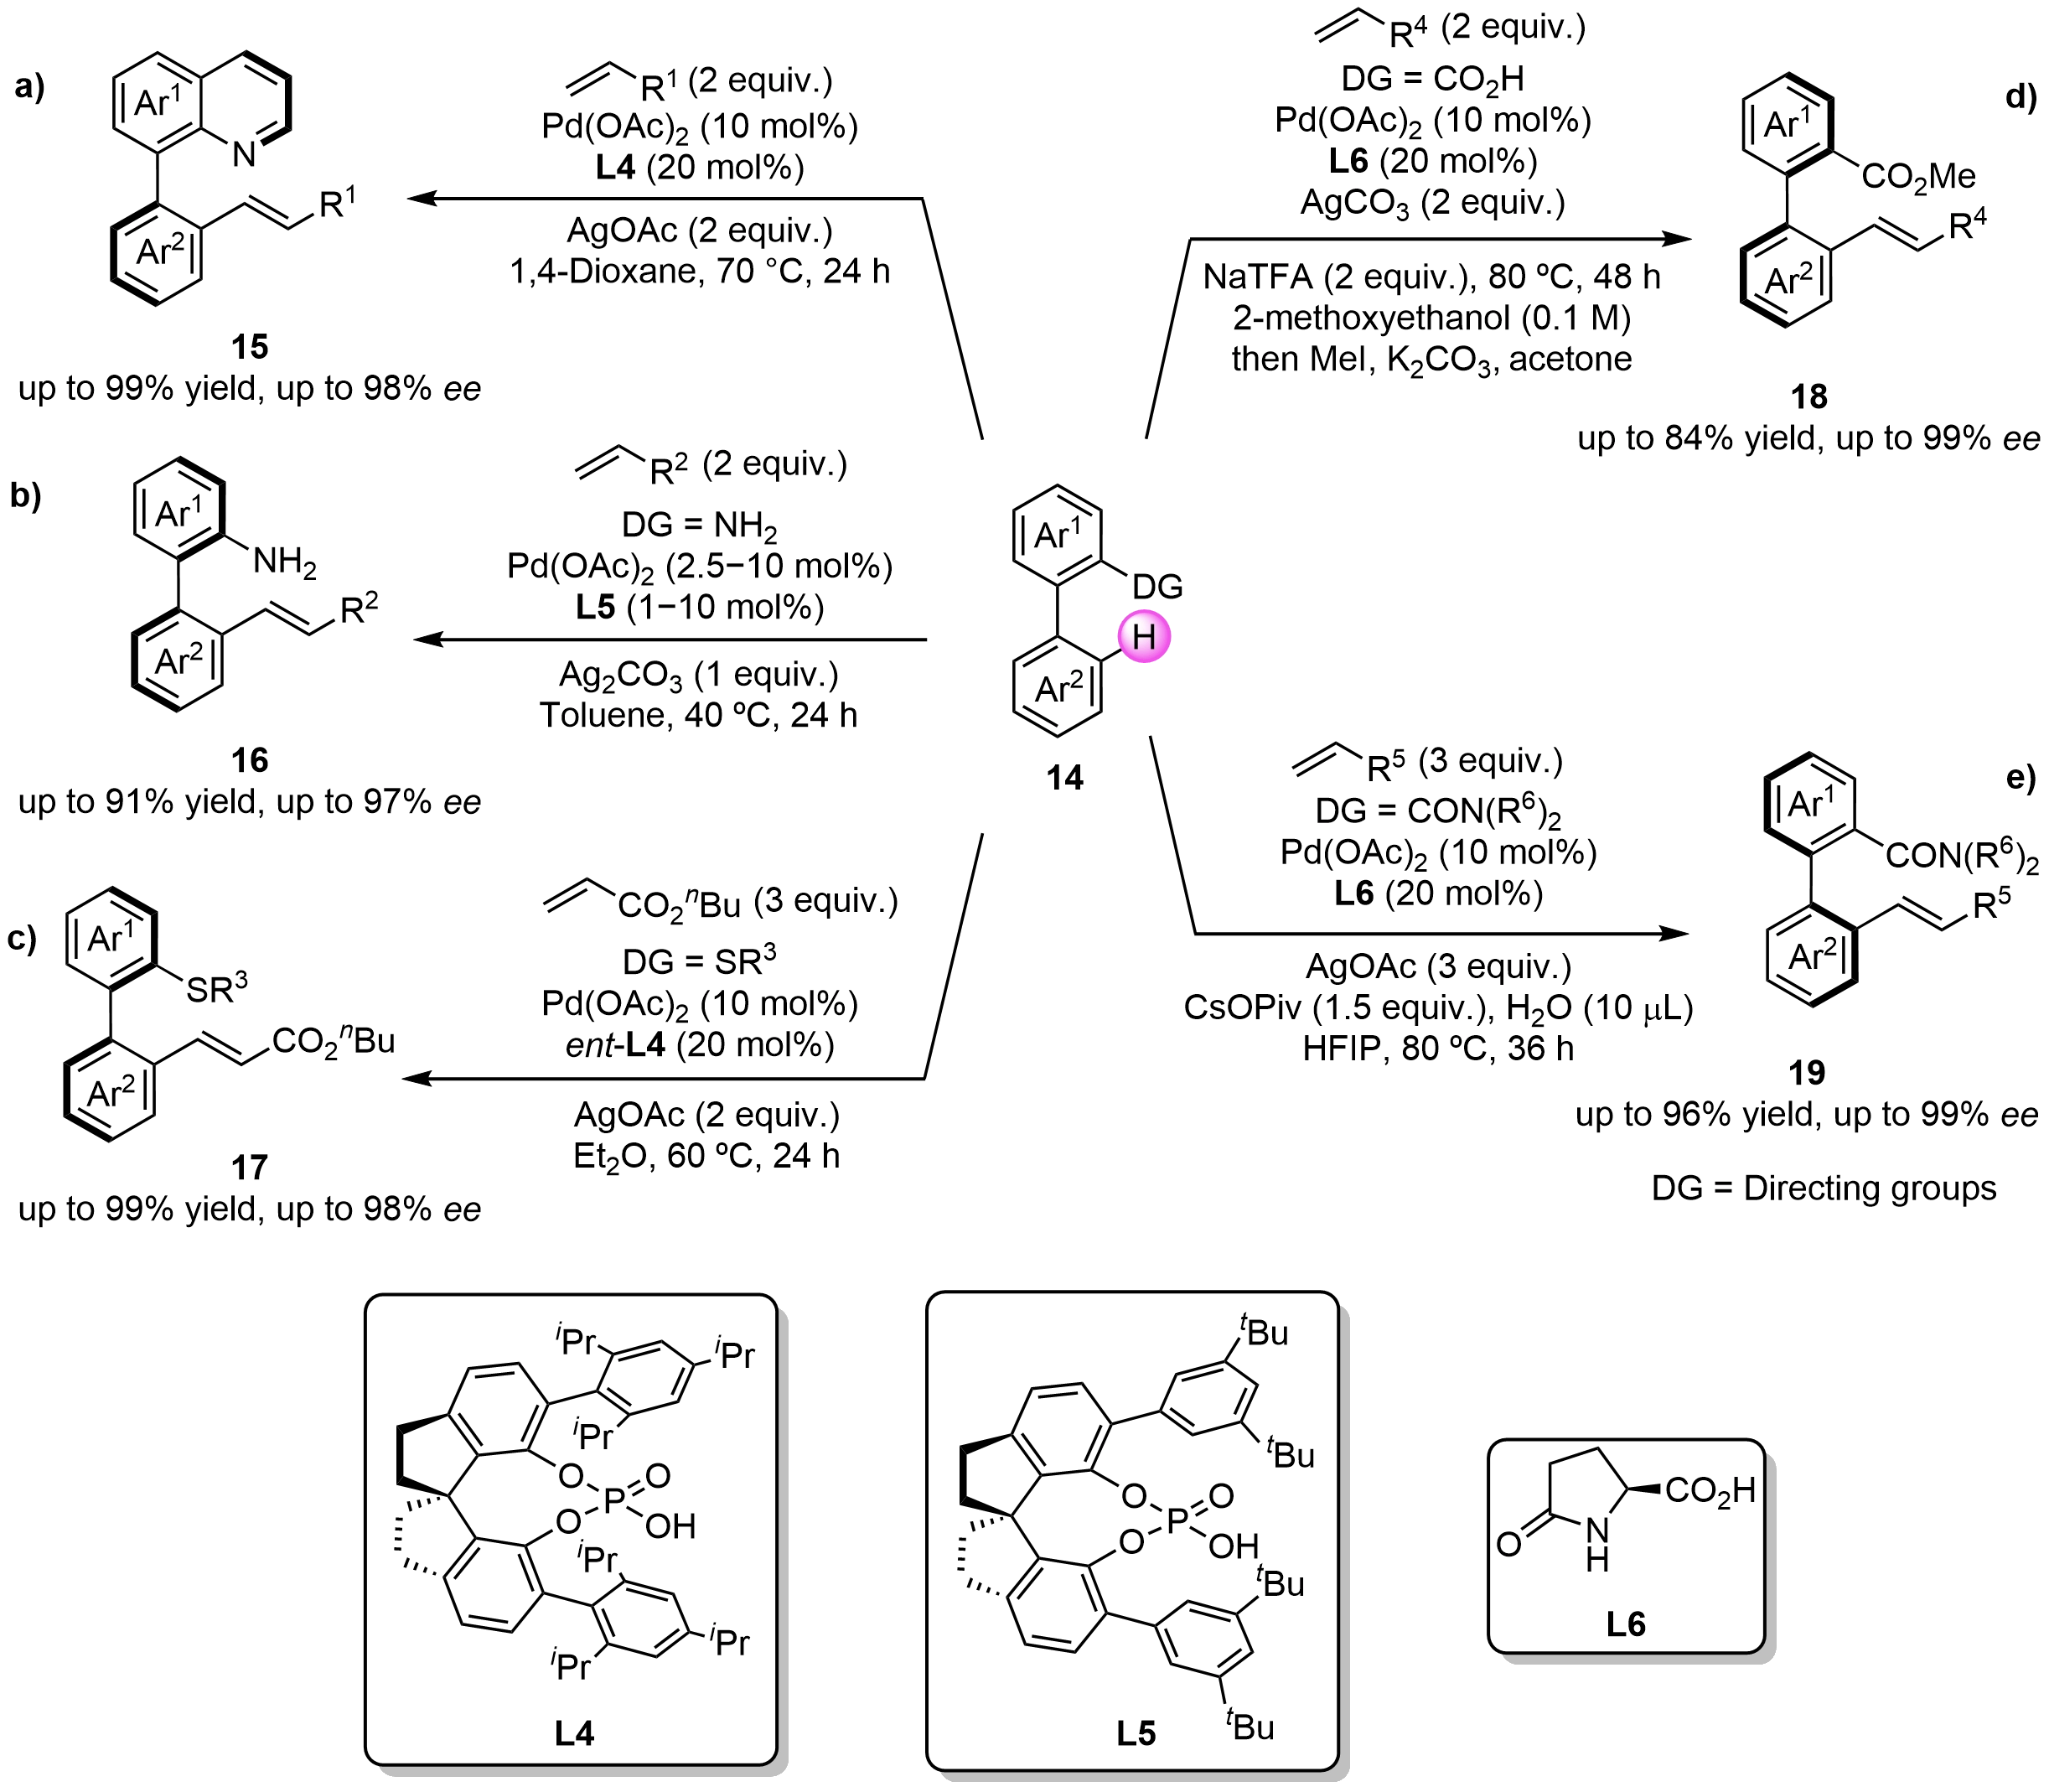
Scheme 9. Construction of C−C axial chirality by palladium-catalyzed asymmetric C−H Heck reactions. DG: directing group.
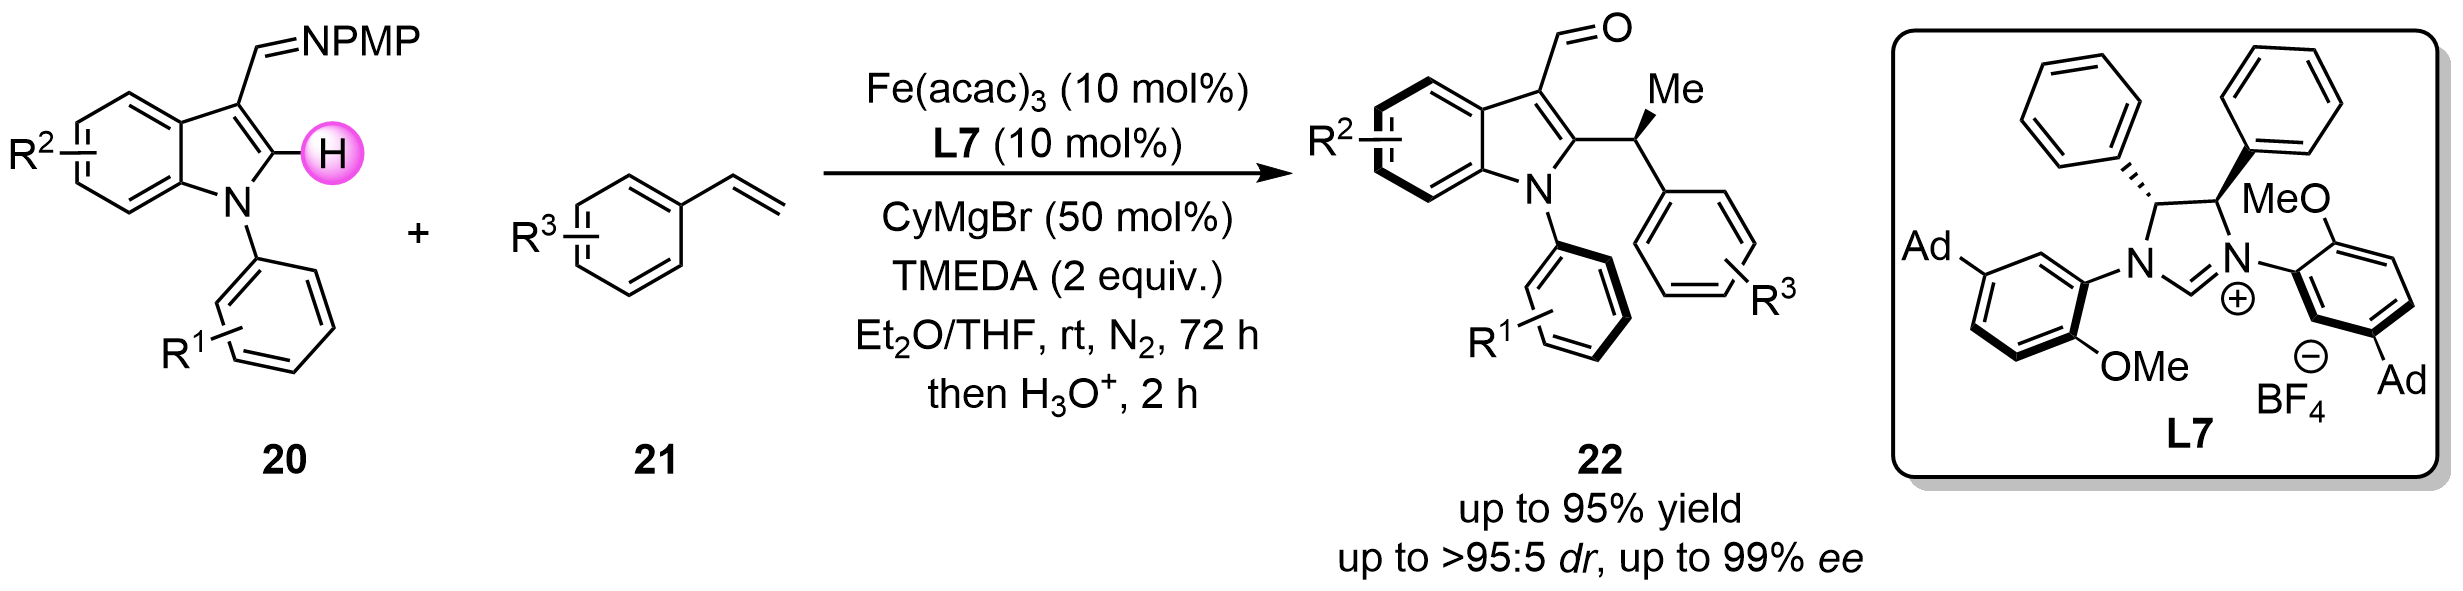
Directing groups can also be utilized in the construction of C−N axial chirality. Compared with six-membered (hetero)biaryl atropisomers, five-membered axially chiral aryl-indole or aryl-pyrrole skeletons exhibit relatively lower rotational barriers, leading to reduced configurational stability[55-57]. In 2024, Ackermann and co-workers described the cooperation between an iron catalyst and an optically active bulky N-heterocyclic carbene (NHC) ligand L7 to achieve C−H alkylation of 1-biarylindoles. Both axial and central chirality could be well constructed in a one-pot operation, and the ligand-to-ligand hydrogen transfer (LLHT) mechanism is also proposed to elaborate this process instead of the classic C−H bond oxidative addition by DFT calculations (Scheme 10)[58].
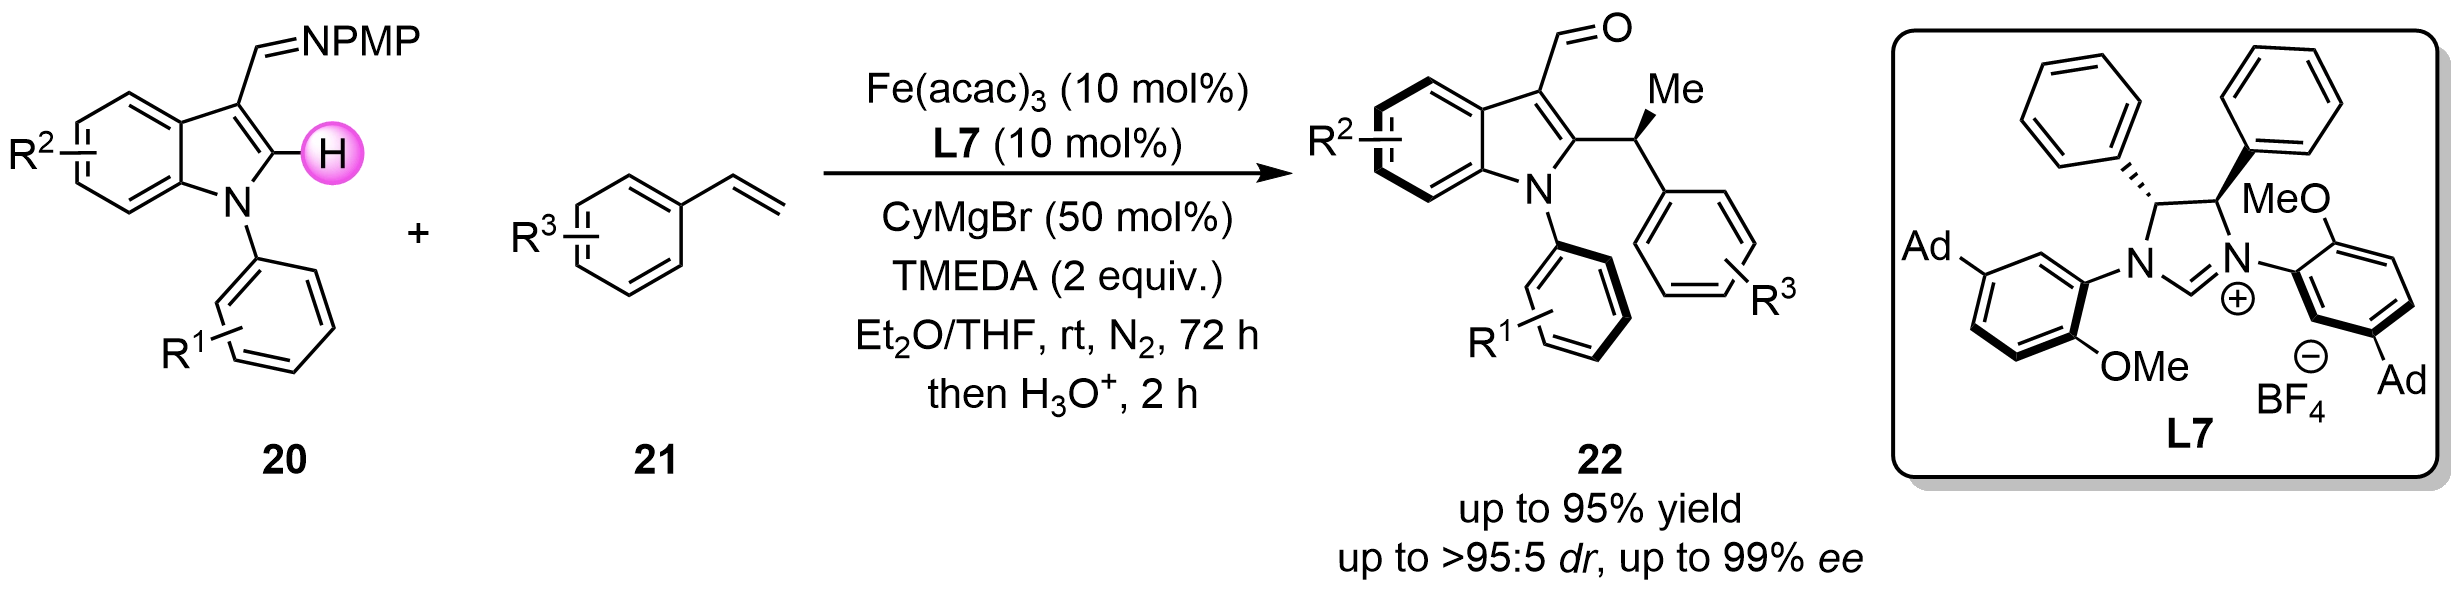
Scheme 10. Construction of C−N axial chirality by iron-catalyzed asymmetric C−H alkylation.
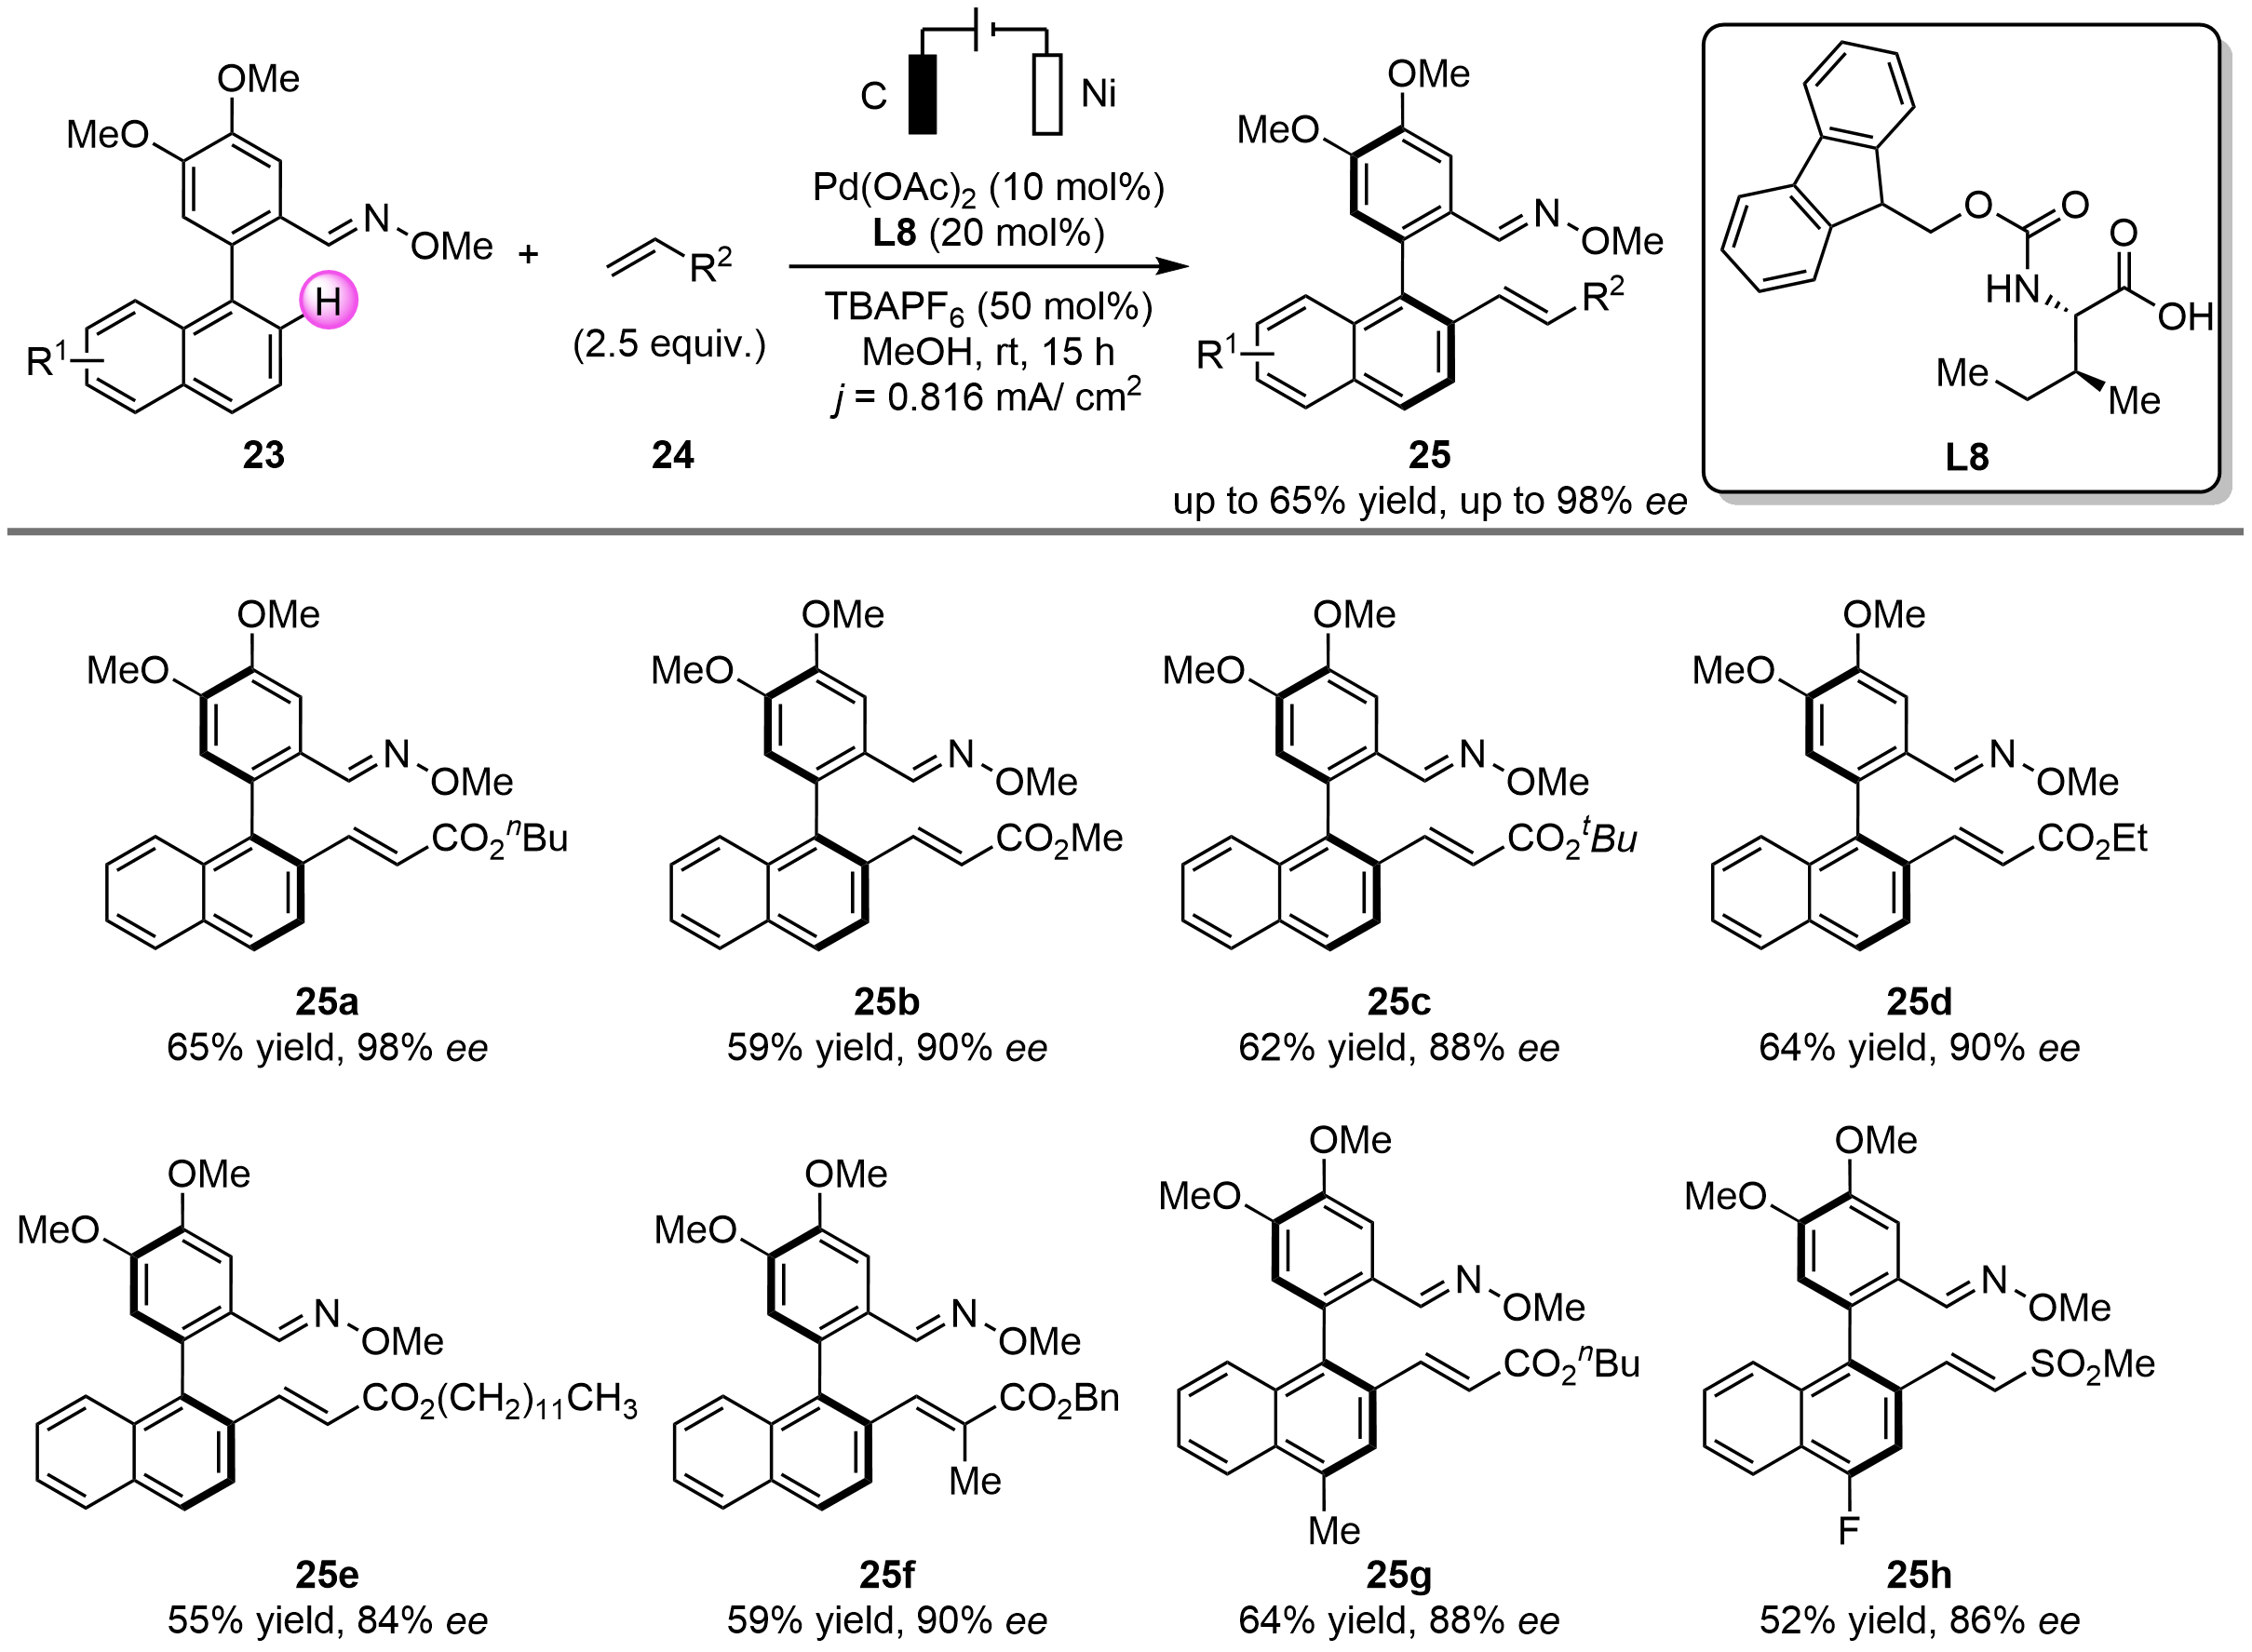
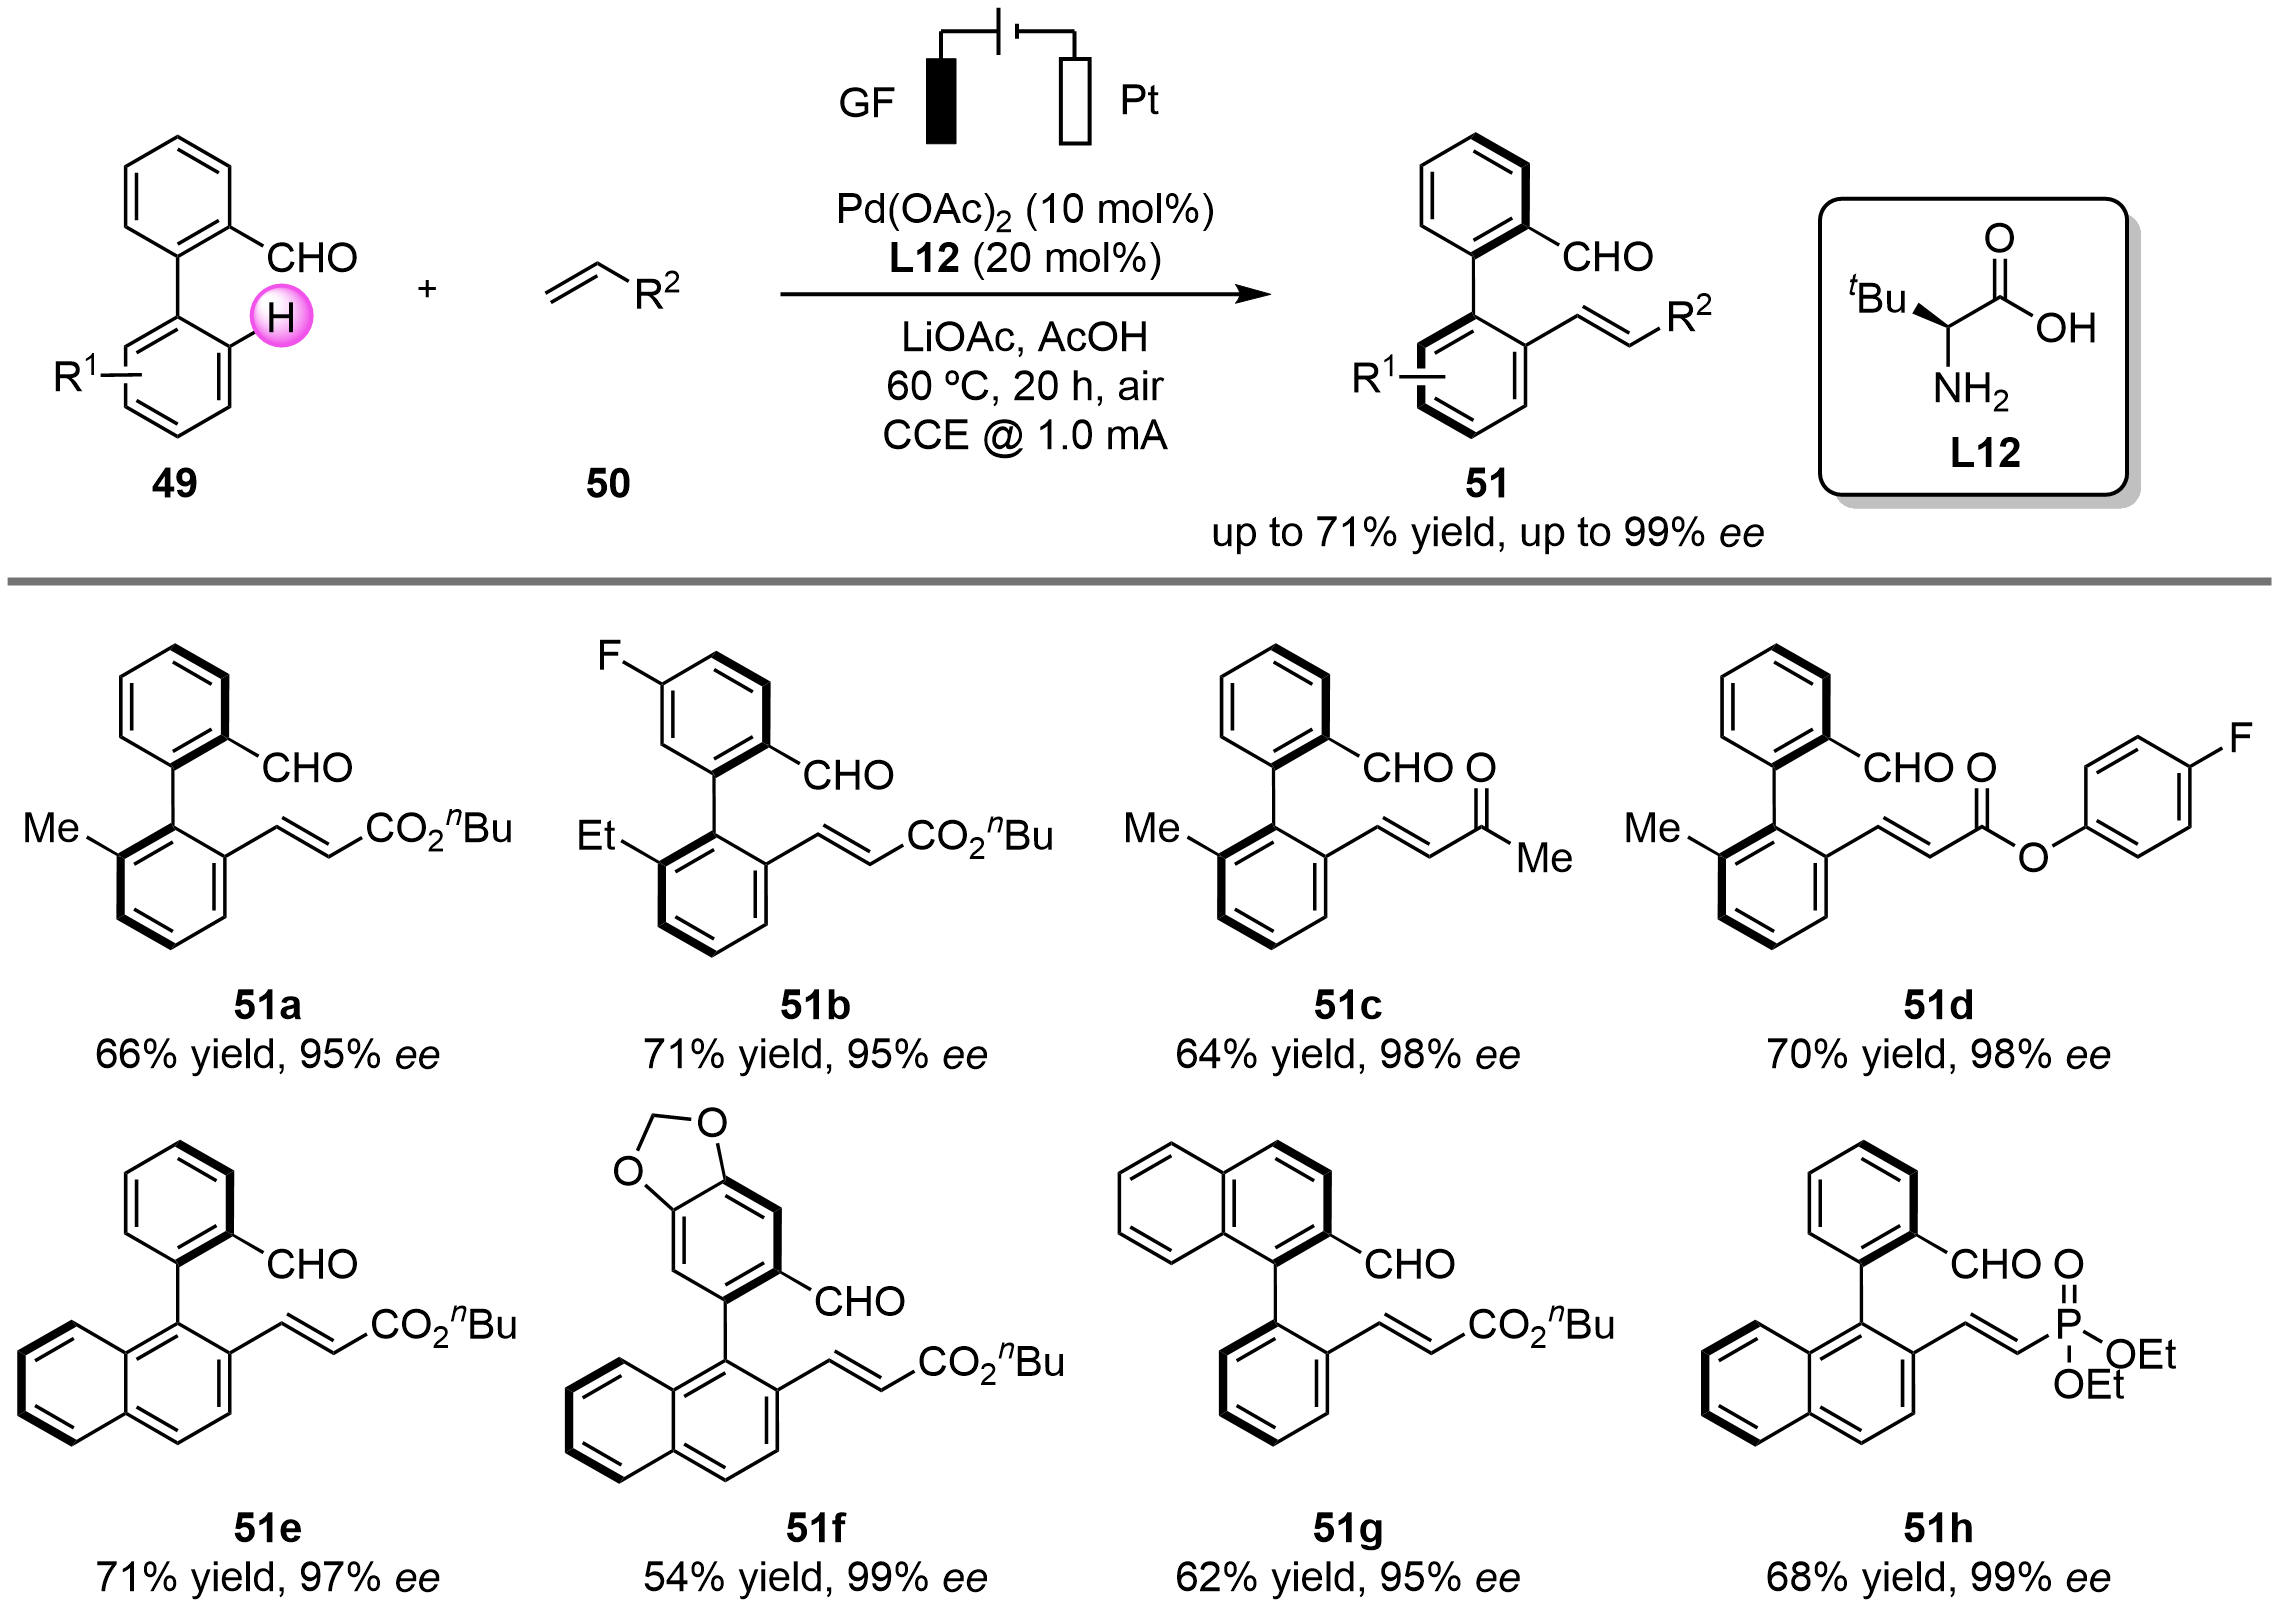
In 2025, the Maiti group reported a Pd-catalyzed C−H olefination of biaryl oximes via an electrochemical pathway. In this transformation, the oxime moiety serves as an effective directing group to enable the electrochemical asymmetric C−H activation. Crucially, the authors noted that the isolated yields for most substrates exceeded 50%, indicating that the reaction proceeds via a DKR process. Furthermore, this strategy offers significant environmental and economic advantages, as it completely obviates the need for traditional metal-based chemical oxidants and external thermal energy (Scheme 11)[59].
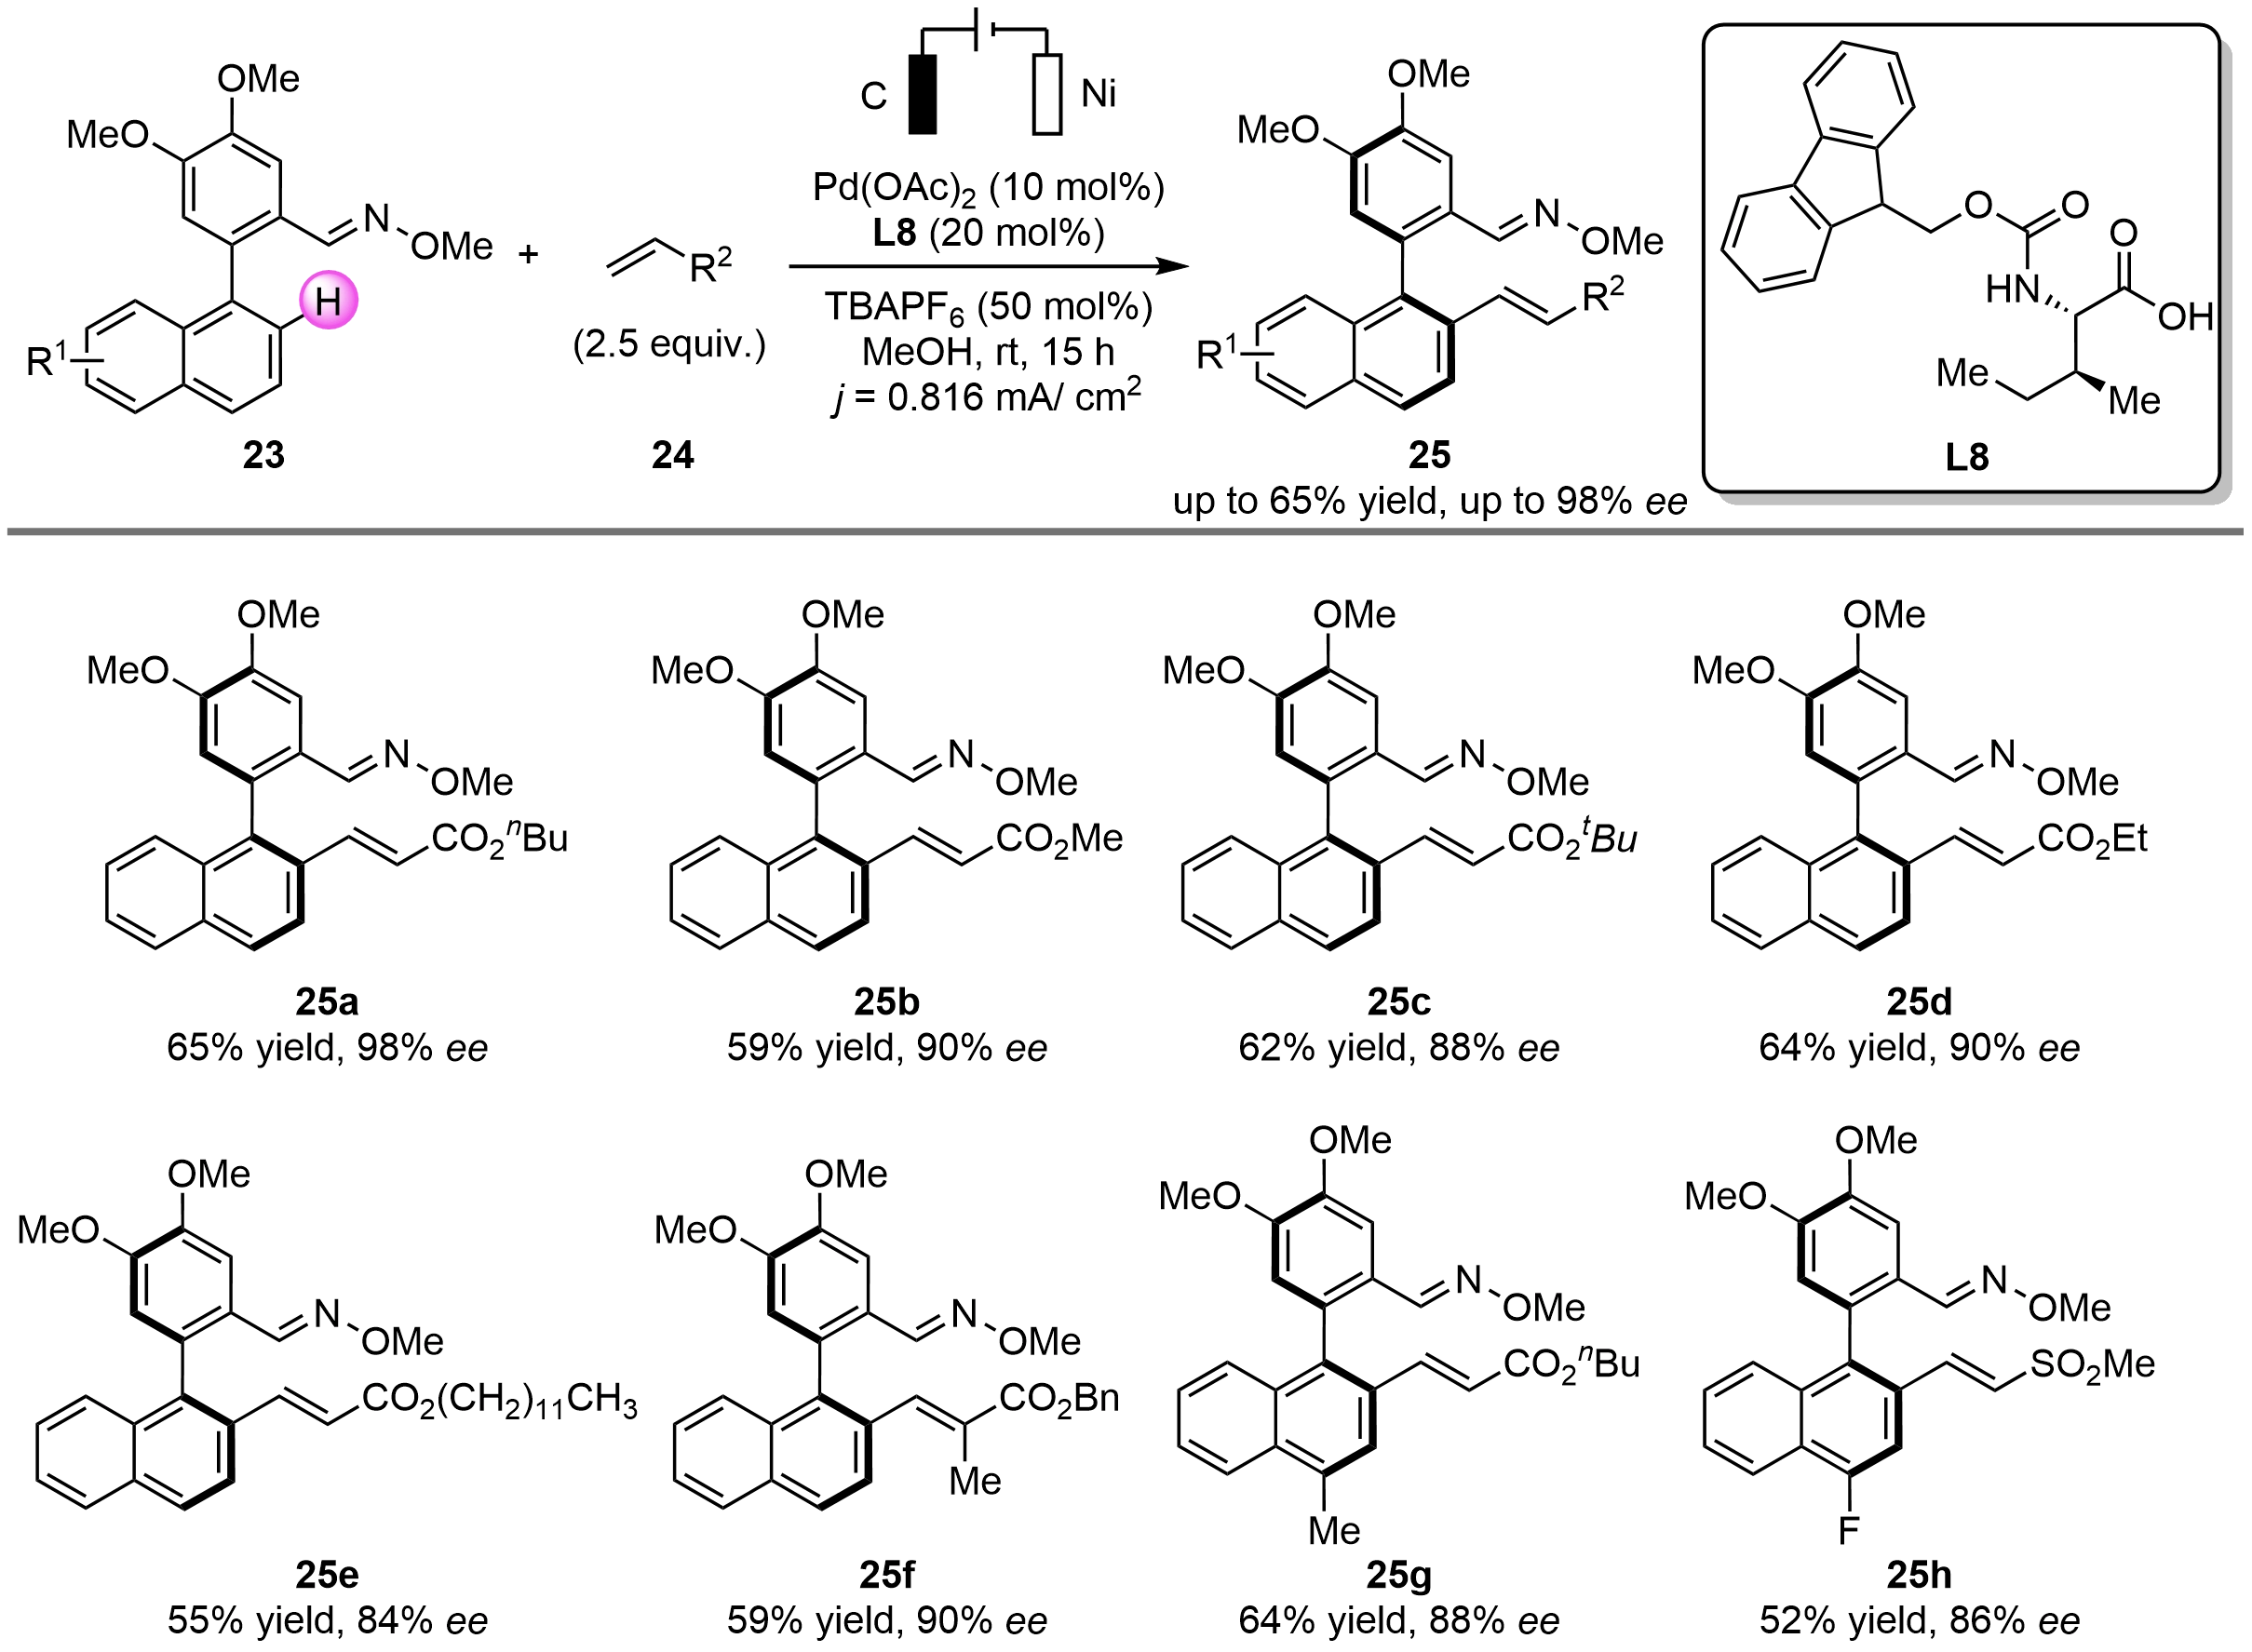
Scheme 11. Construction of C−C axial chirality by palladium-catalyzed electrochemical asymmetric C−H olefination.
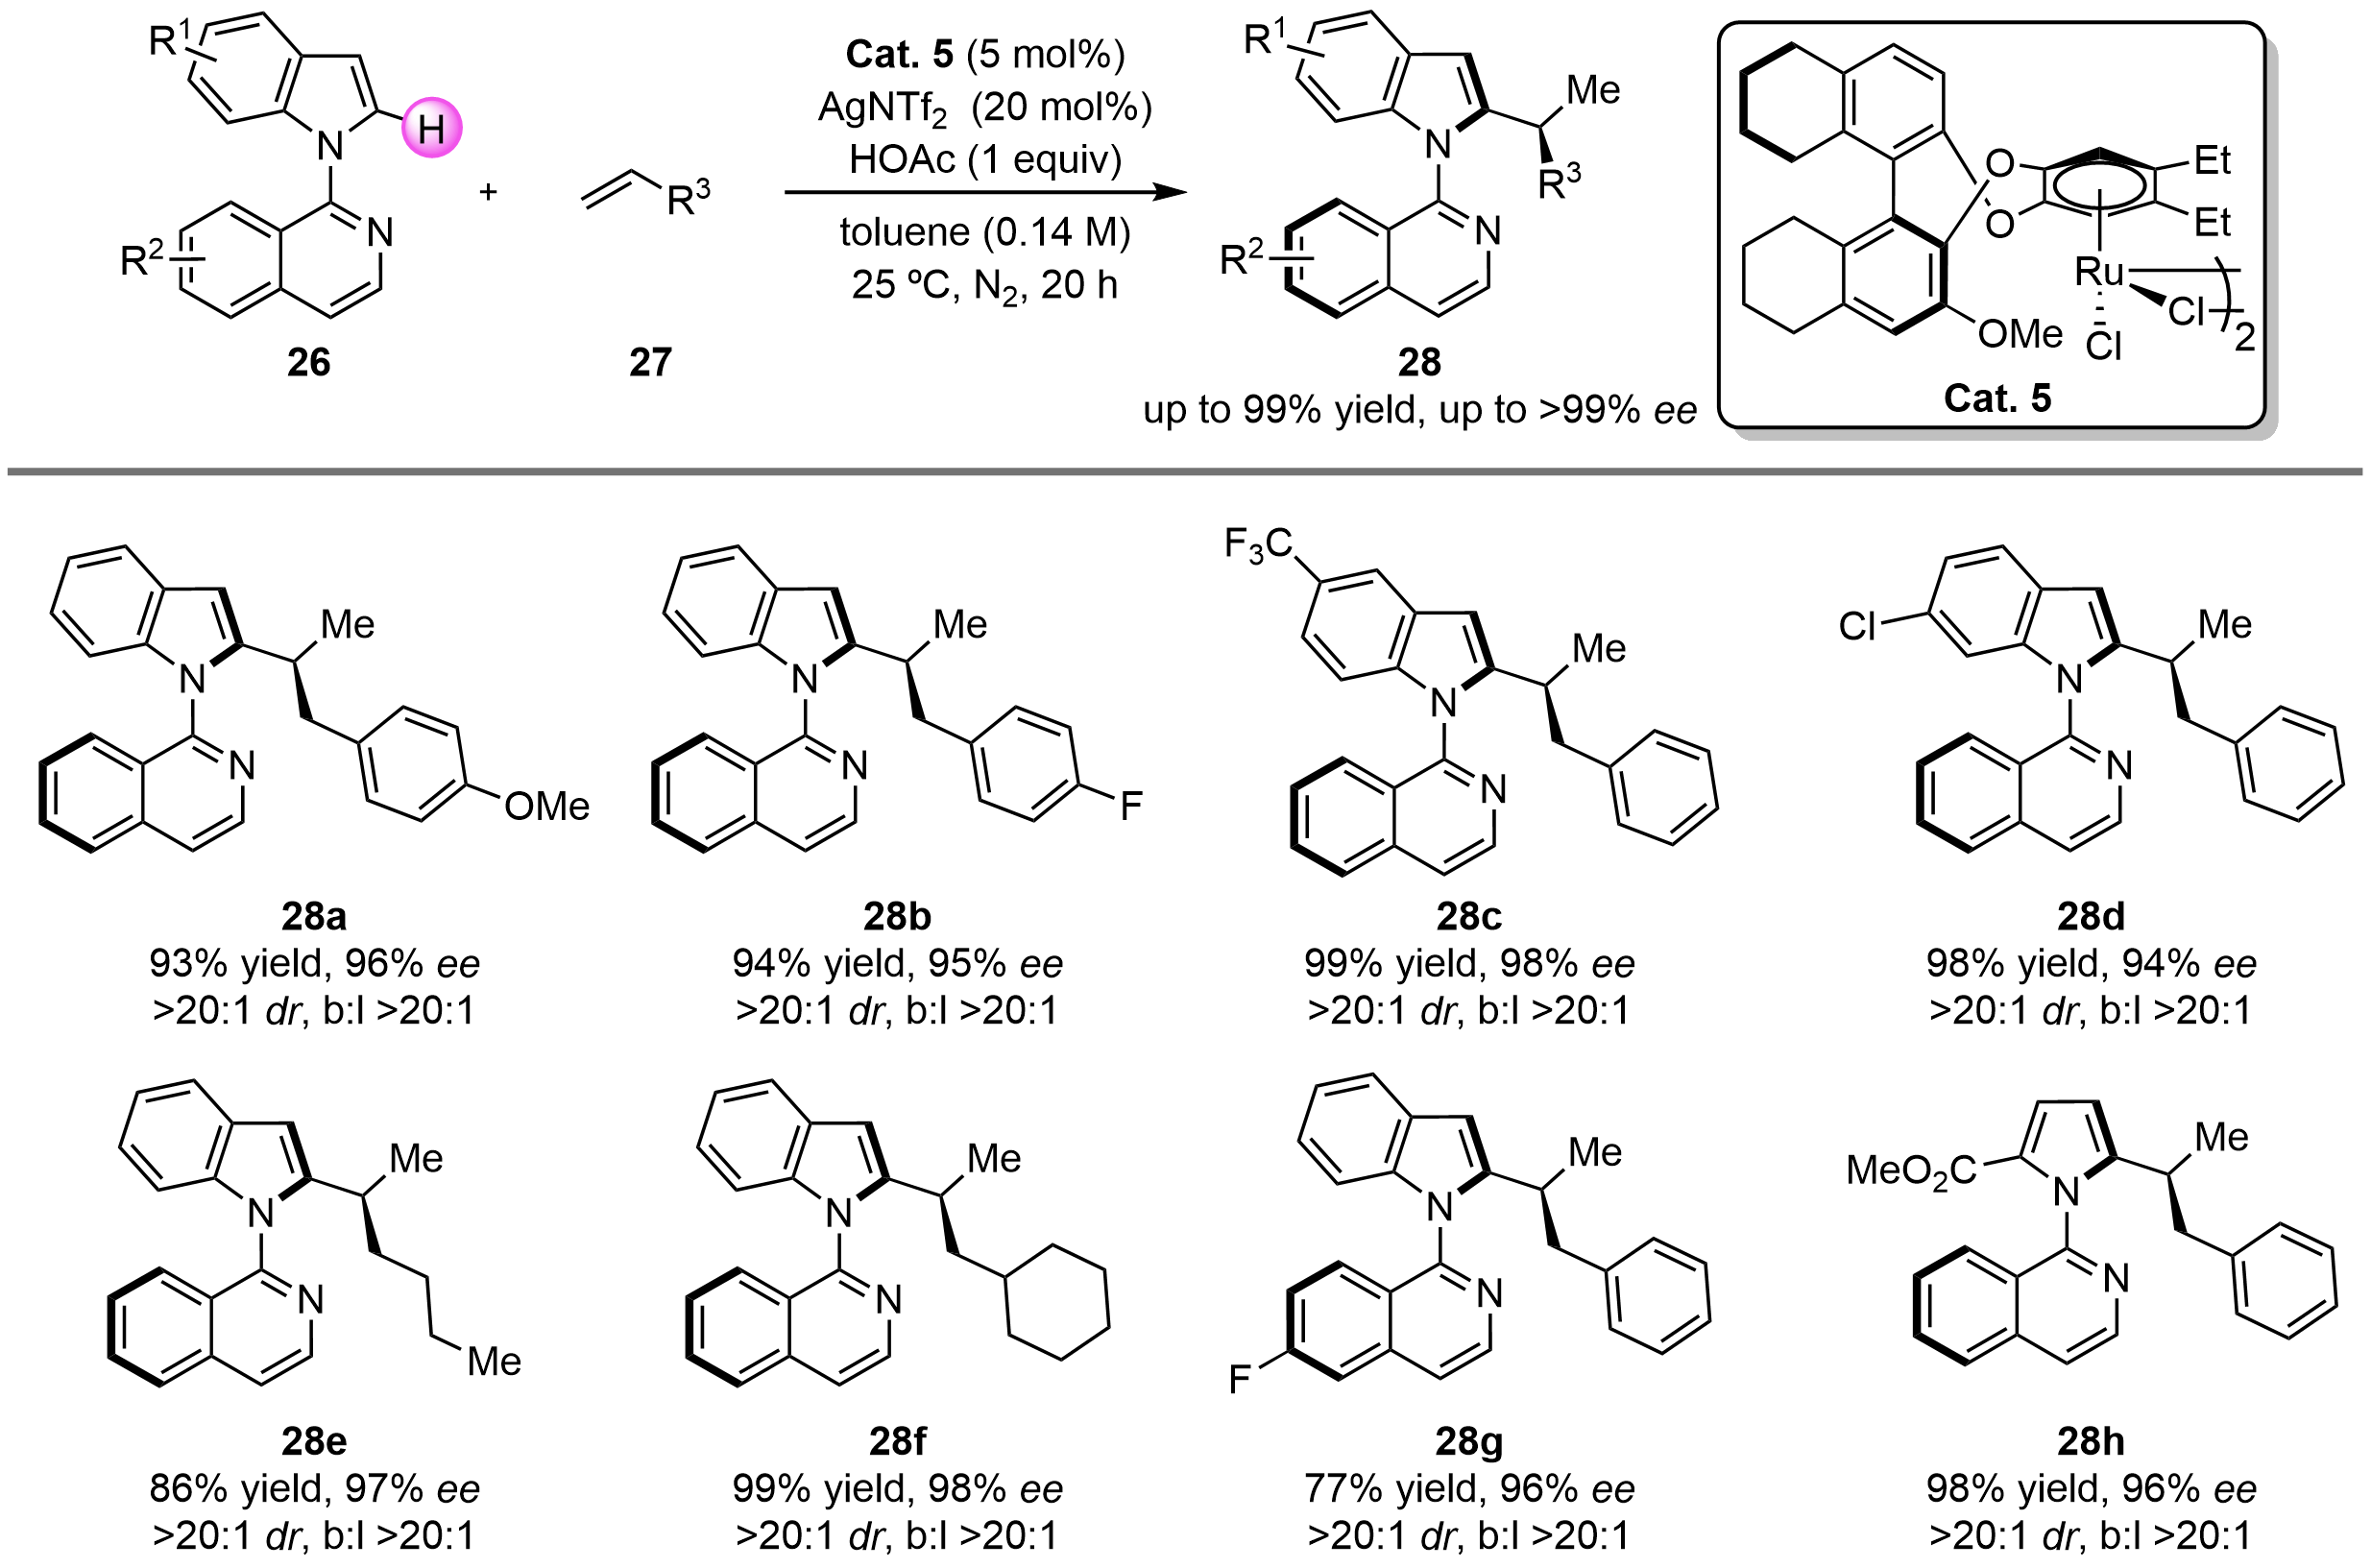
In 2026, the Wang group employed chiral η6-benzene-ruthenium catalysts along with the directing effect of a quinoline moiety to generate a range of chiral indoles bearing both central and axial chirality (Scheme 12)[60]. In this work, the asymmetric C−H alkylation of indoles with unactivated terminal alkenes was carried out, thus simultaneously establishing both central and axial chirality. The reaction exhibited a broad substrate scope and excellent functional-group tolerance.
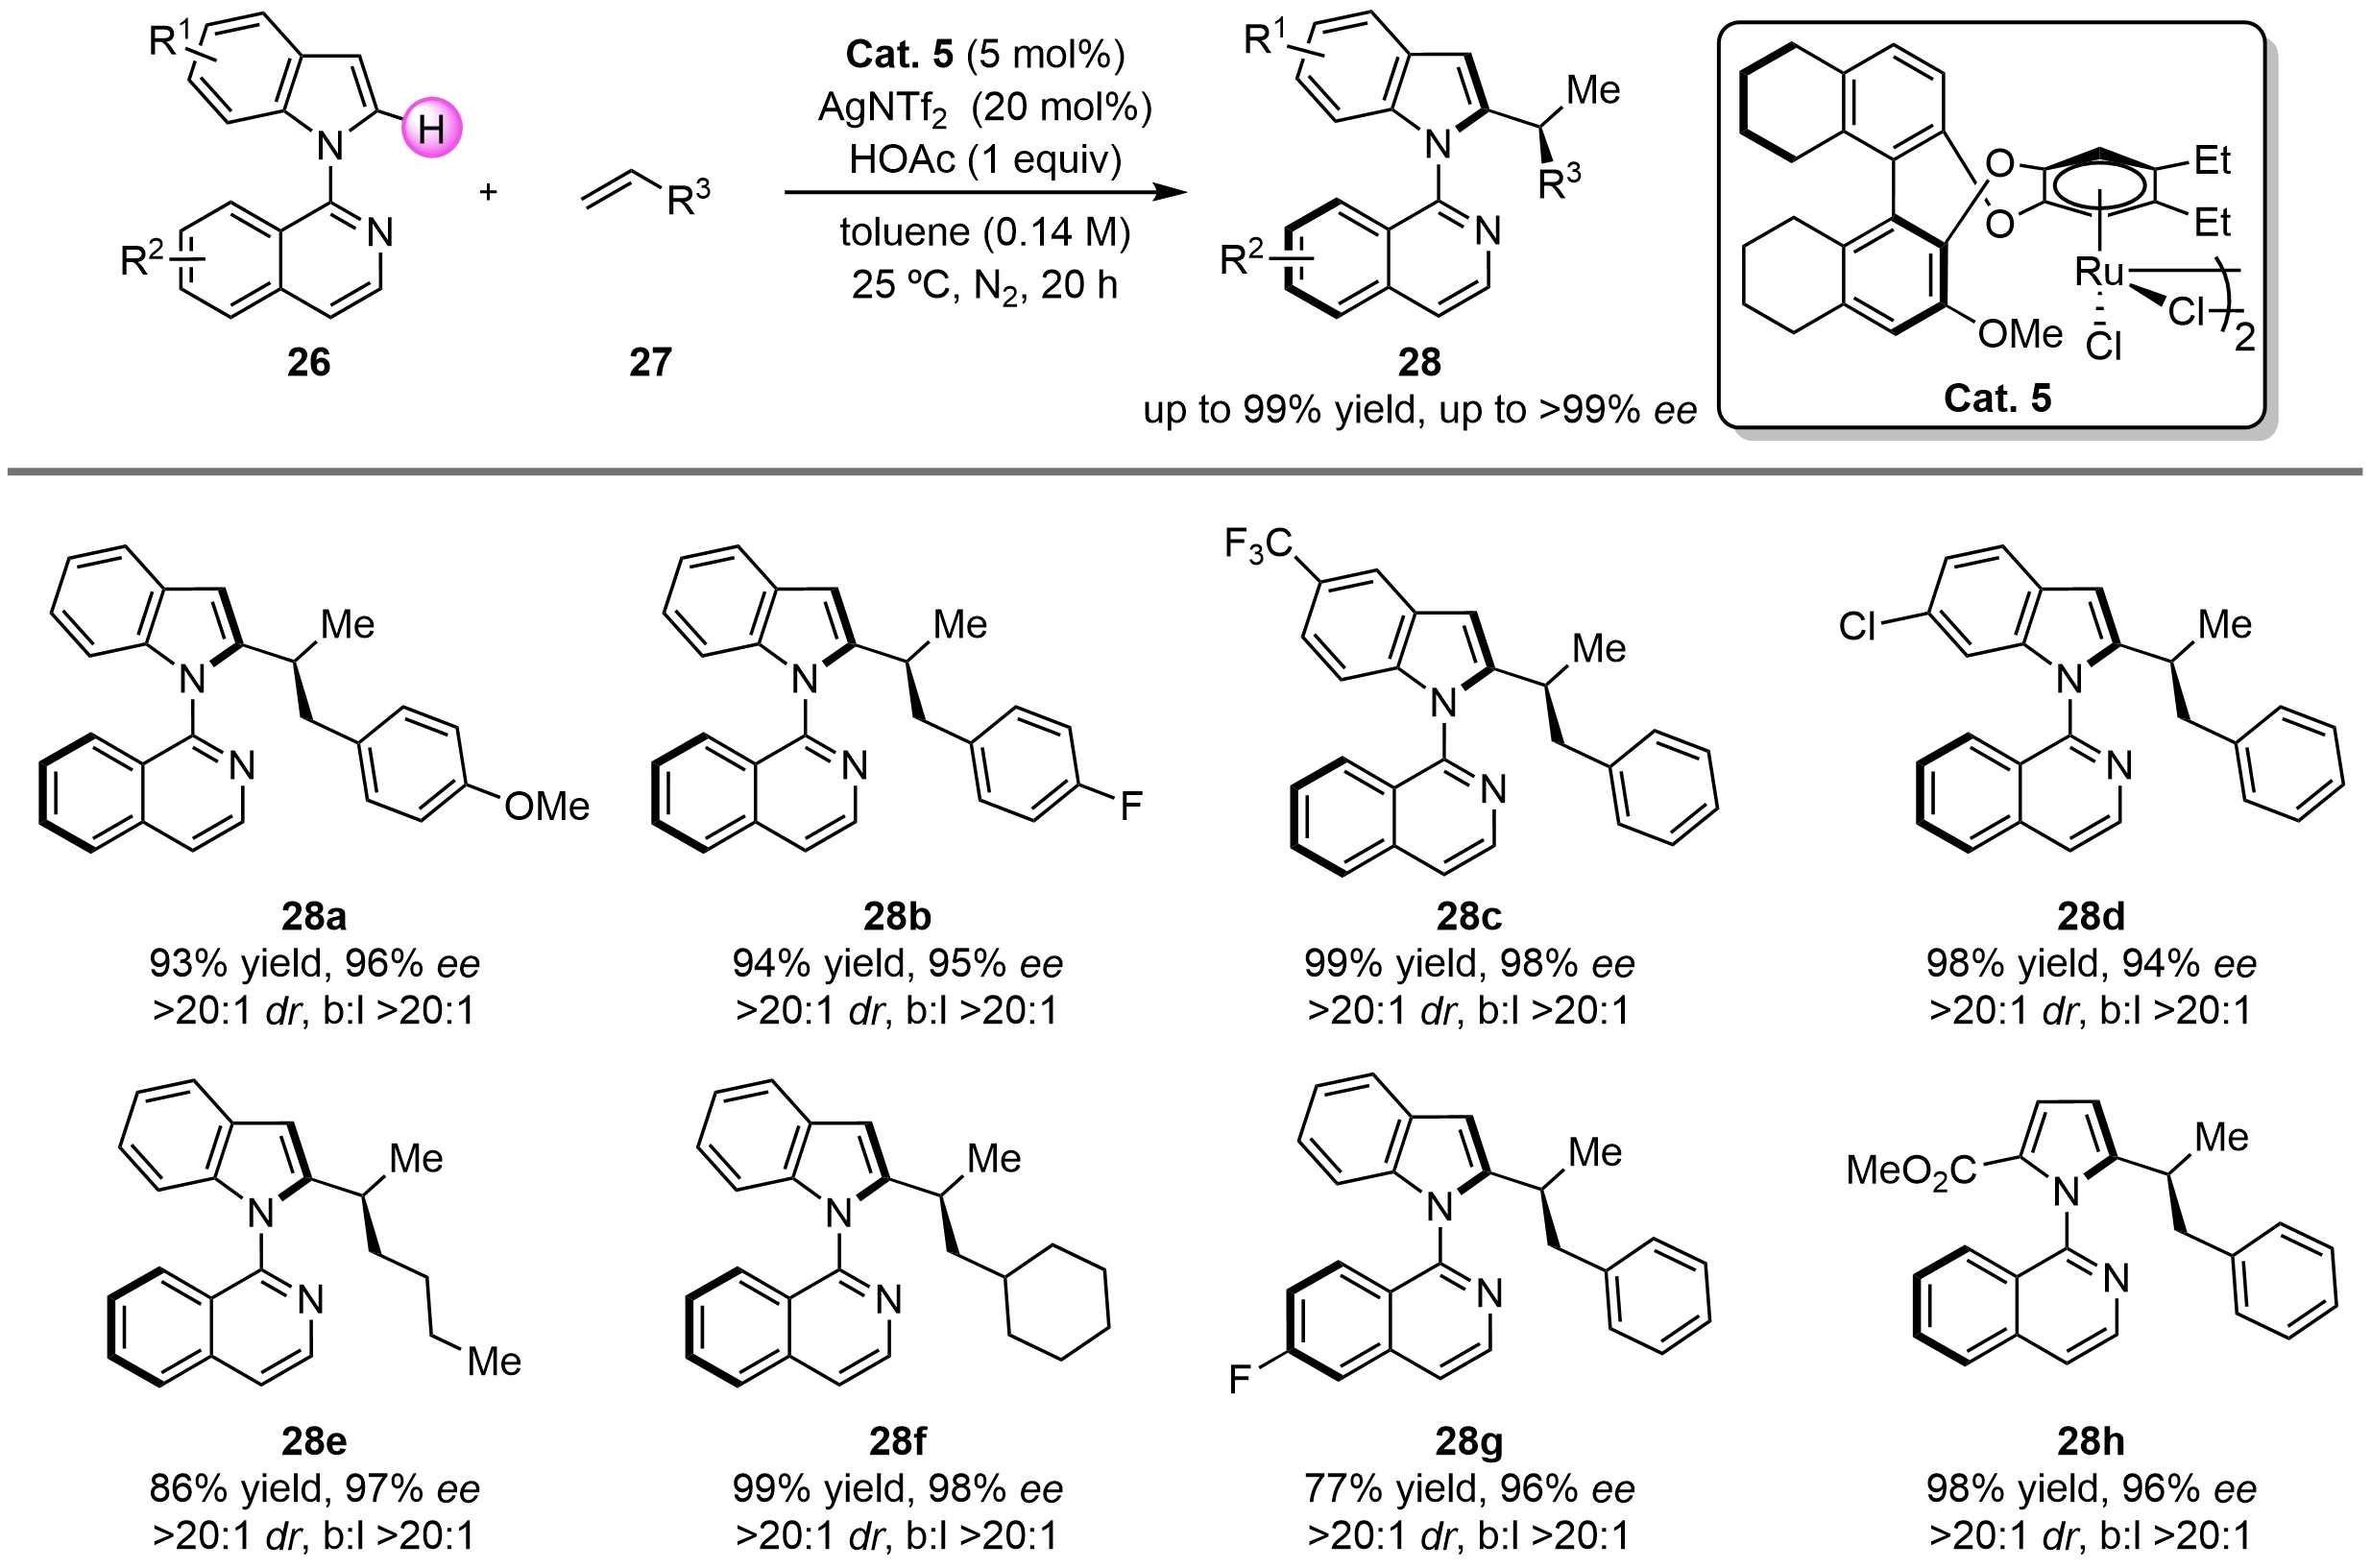
Scheme 12. Construction of C−N axial chirality by ruthenium-catalyzed asymmetric C−H alkylation.
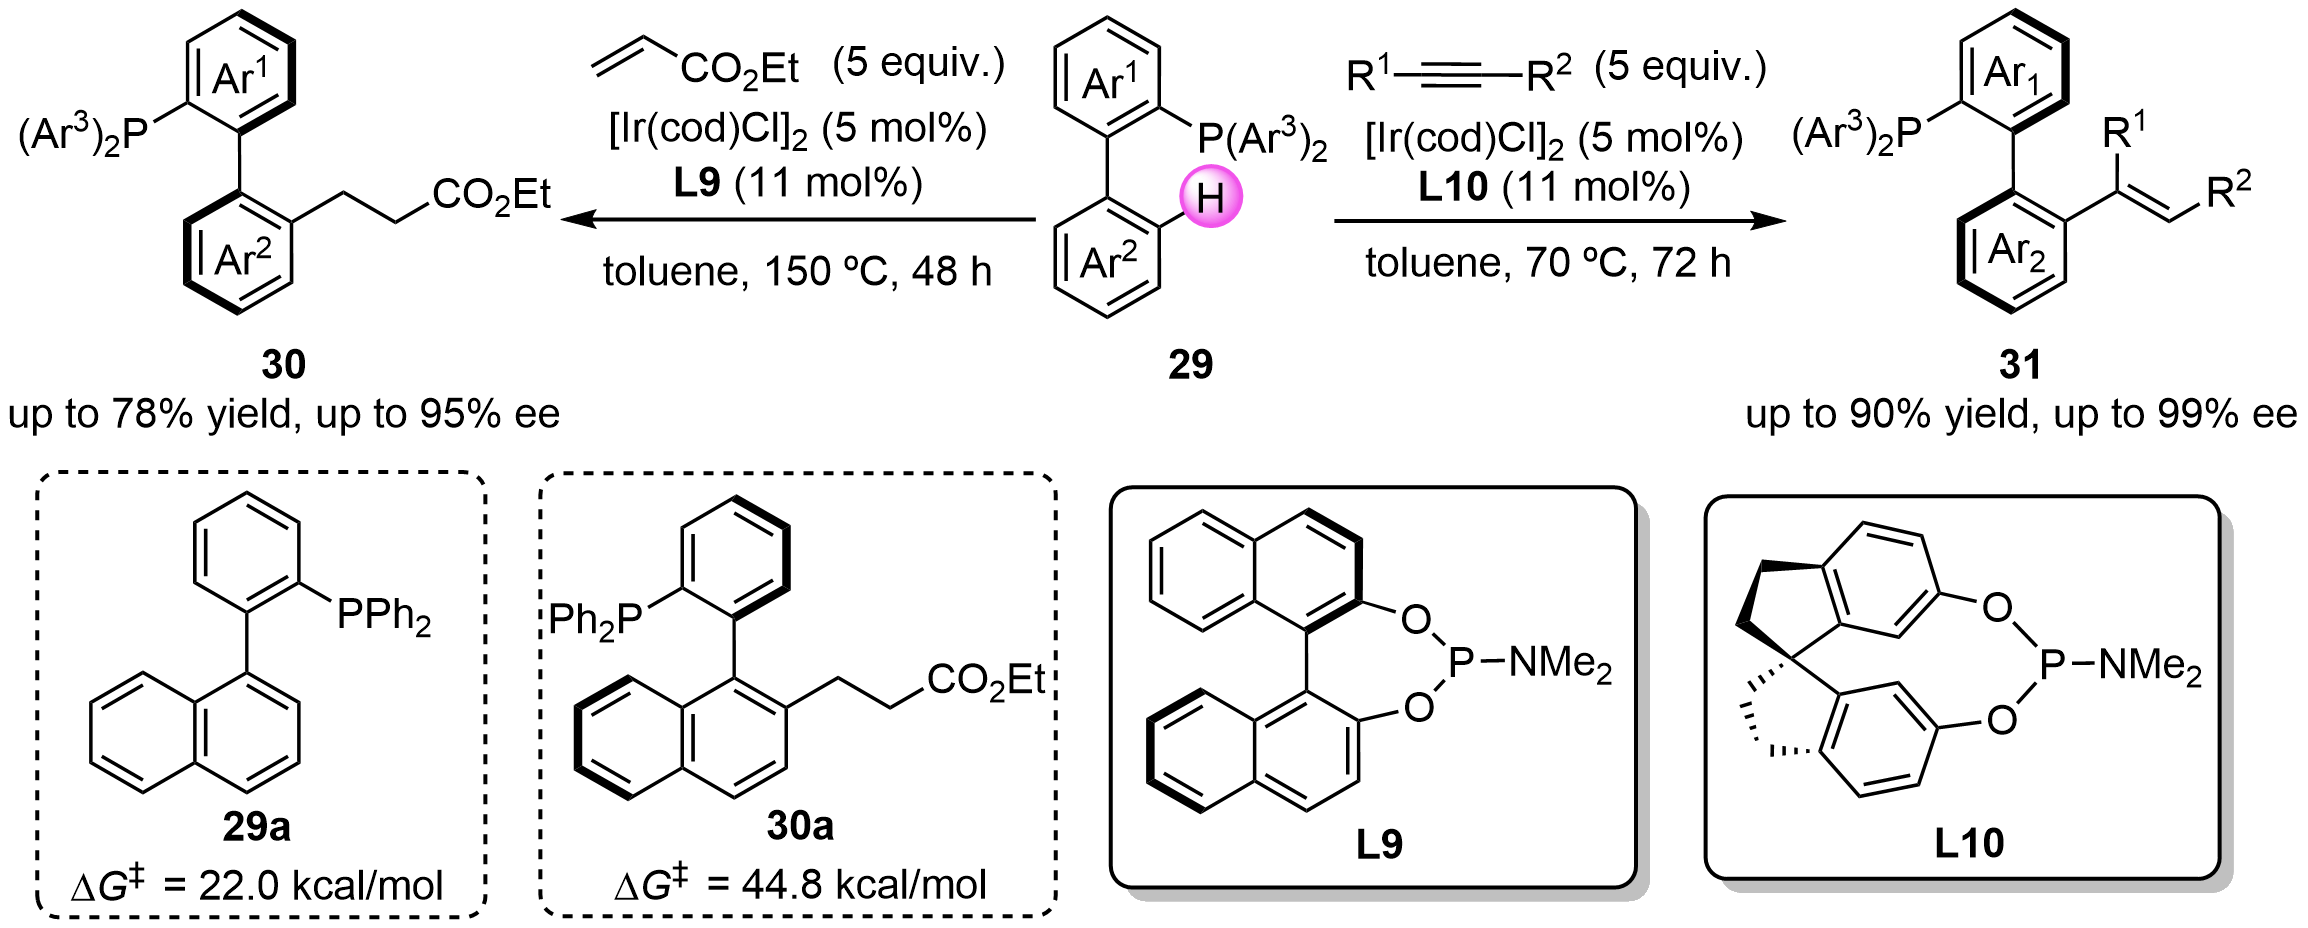
In 2023, the Shi group developed iridium-catalyzed asymmetric C−H olefination and alkylation reactions of biaryl phosphines directed by P(III) centers. Under the optimized conditions, a broad range of biaryl phosphines underwent the hydroarylation of alkynes or alkenes smoothly via a DKR pathway, affording a library of axially chiral functionalized phosphines in high yields with exceptional enantioselectivities. Crucially, the calculated rotational barrier for the starting biaryl phosphine substrate 29a is 22.0 kcal/mol, indicating facile racemization. In stark contrast, the inversion barrier of the functionalized product 30a sharply increases to 44.8 kcal/mol, rendering it configurationally stable and highly resistant to racemization. Consequently, these energetic profiles substantiate that a successful DKR is effectively achieved through this phosphorus-directed C−H activation (Scheme 13)[61]. It is worth noting that the synthetic utility of these generated axially chiral biaryl phosphines was successfully demonstrated by their use as highly efficient ligands in rhodium- and palladium-catalyzed asymmetric transformations.
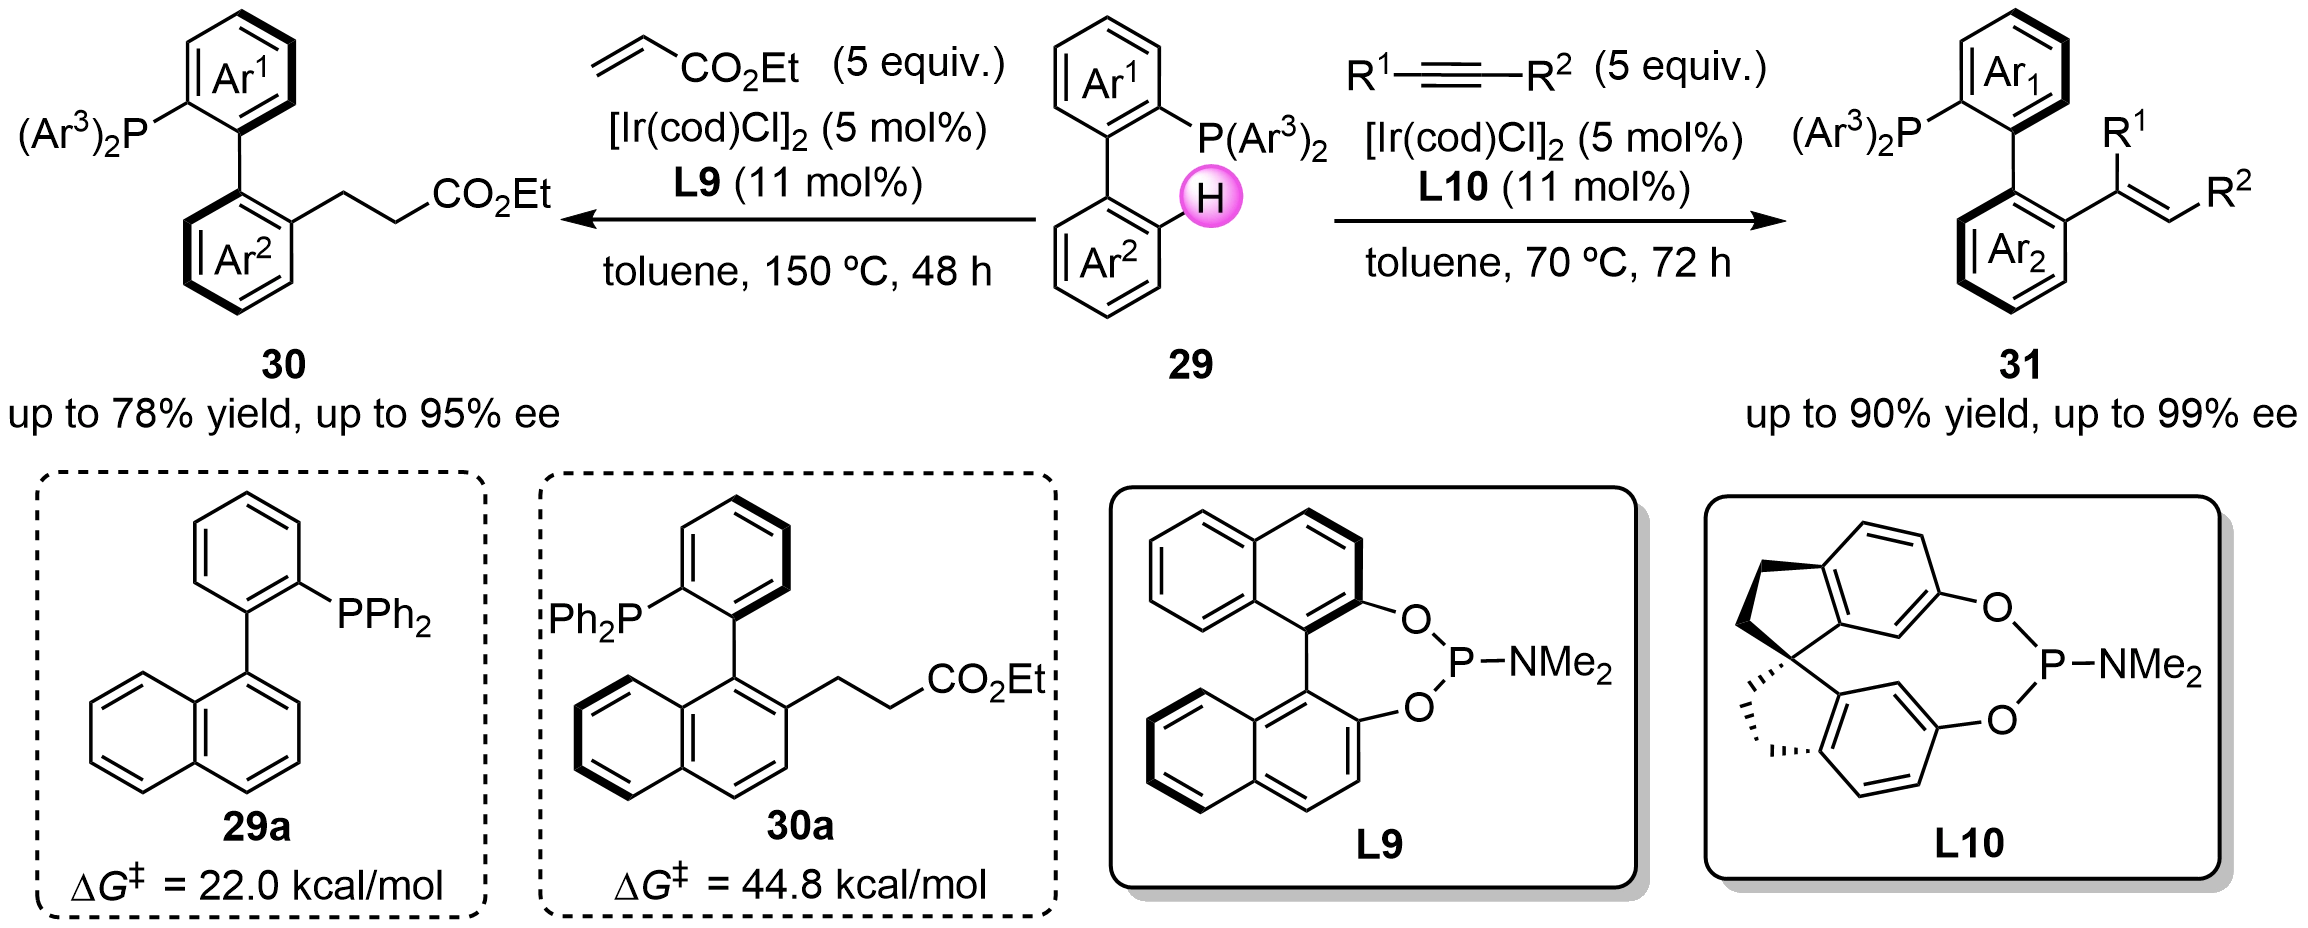
Scheme 13. Construction of C−C axial chirality by iridium-catalyzed asymmetric C−H olefination and alkylation.
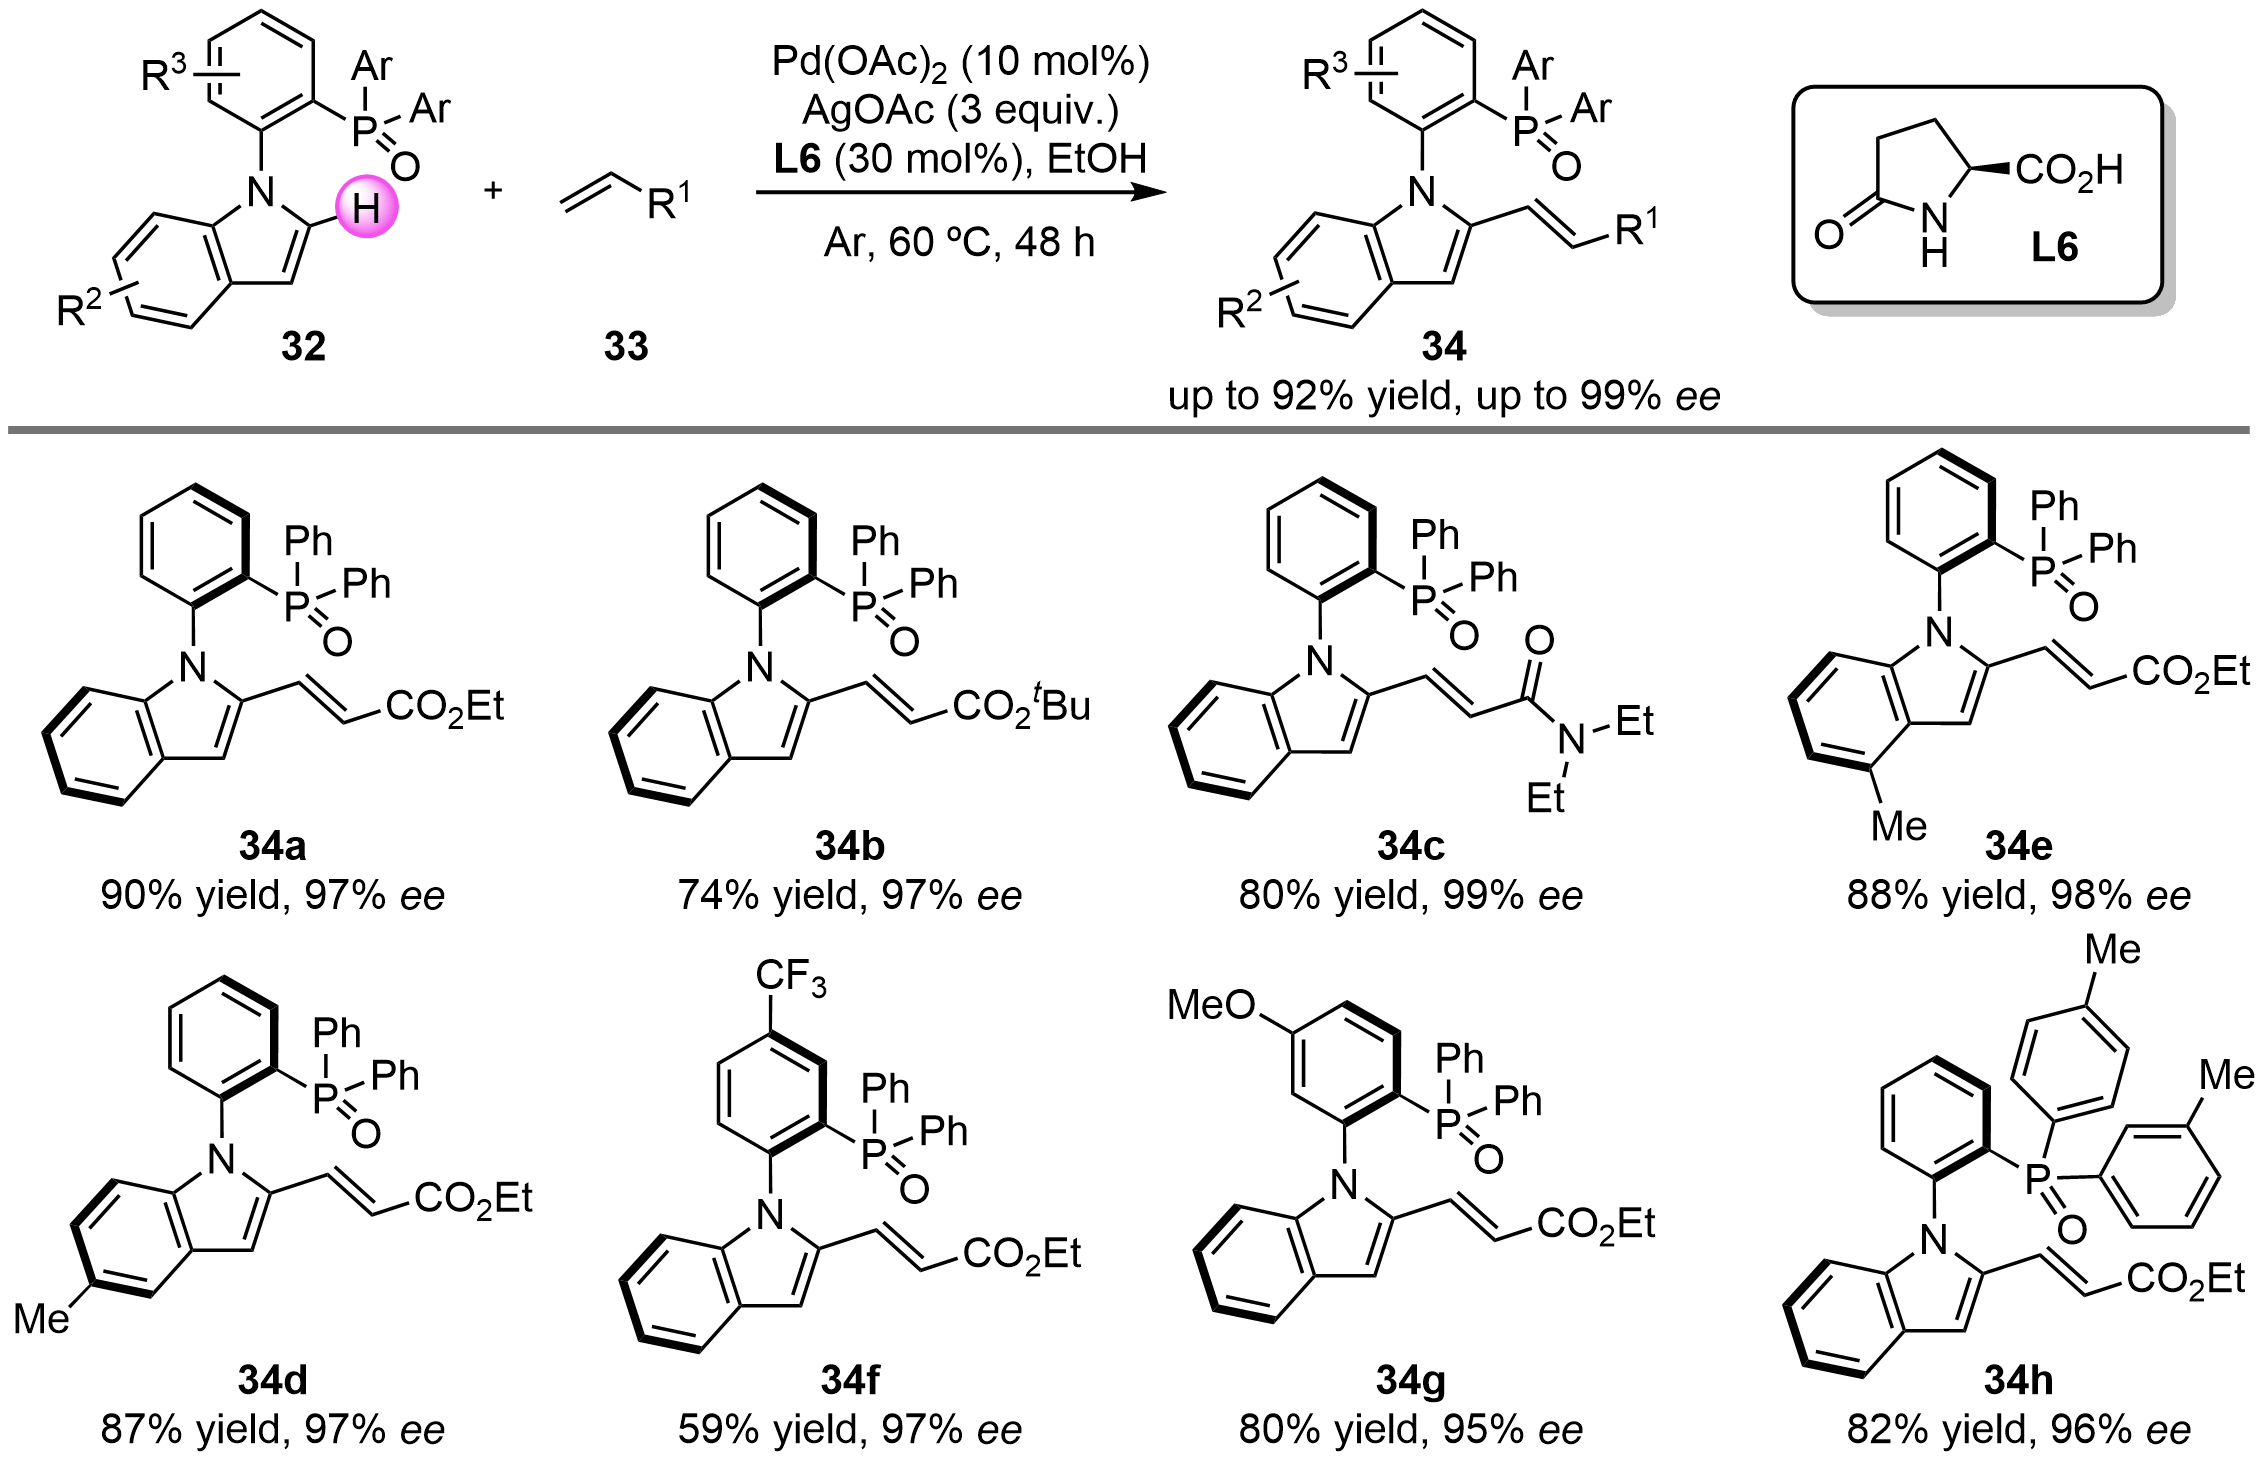
In 2025, the Li group developed a novel P(=O)Ar2-directed asymmetric catalytic olefination. This work represents the first instance of installing a P(=O)Ar2 group at the ortho-position of the N-aryl moiety of indoles as a directing group, successfully achieving an asymmetric olefination reaction of C−H bonds and constructing C−N axial chirality (Scheme 14)[62].
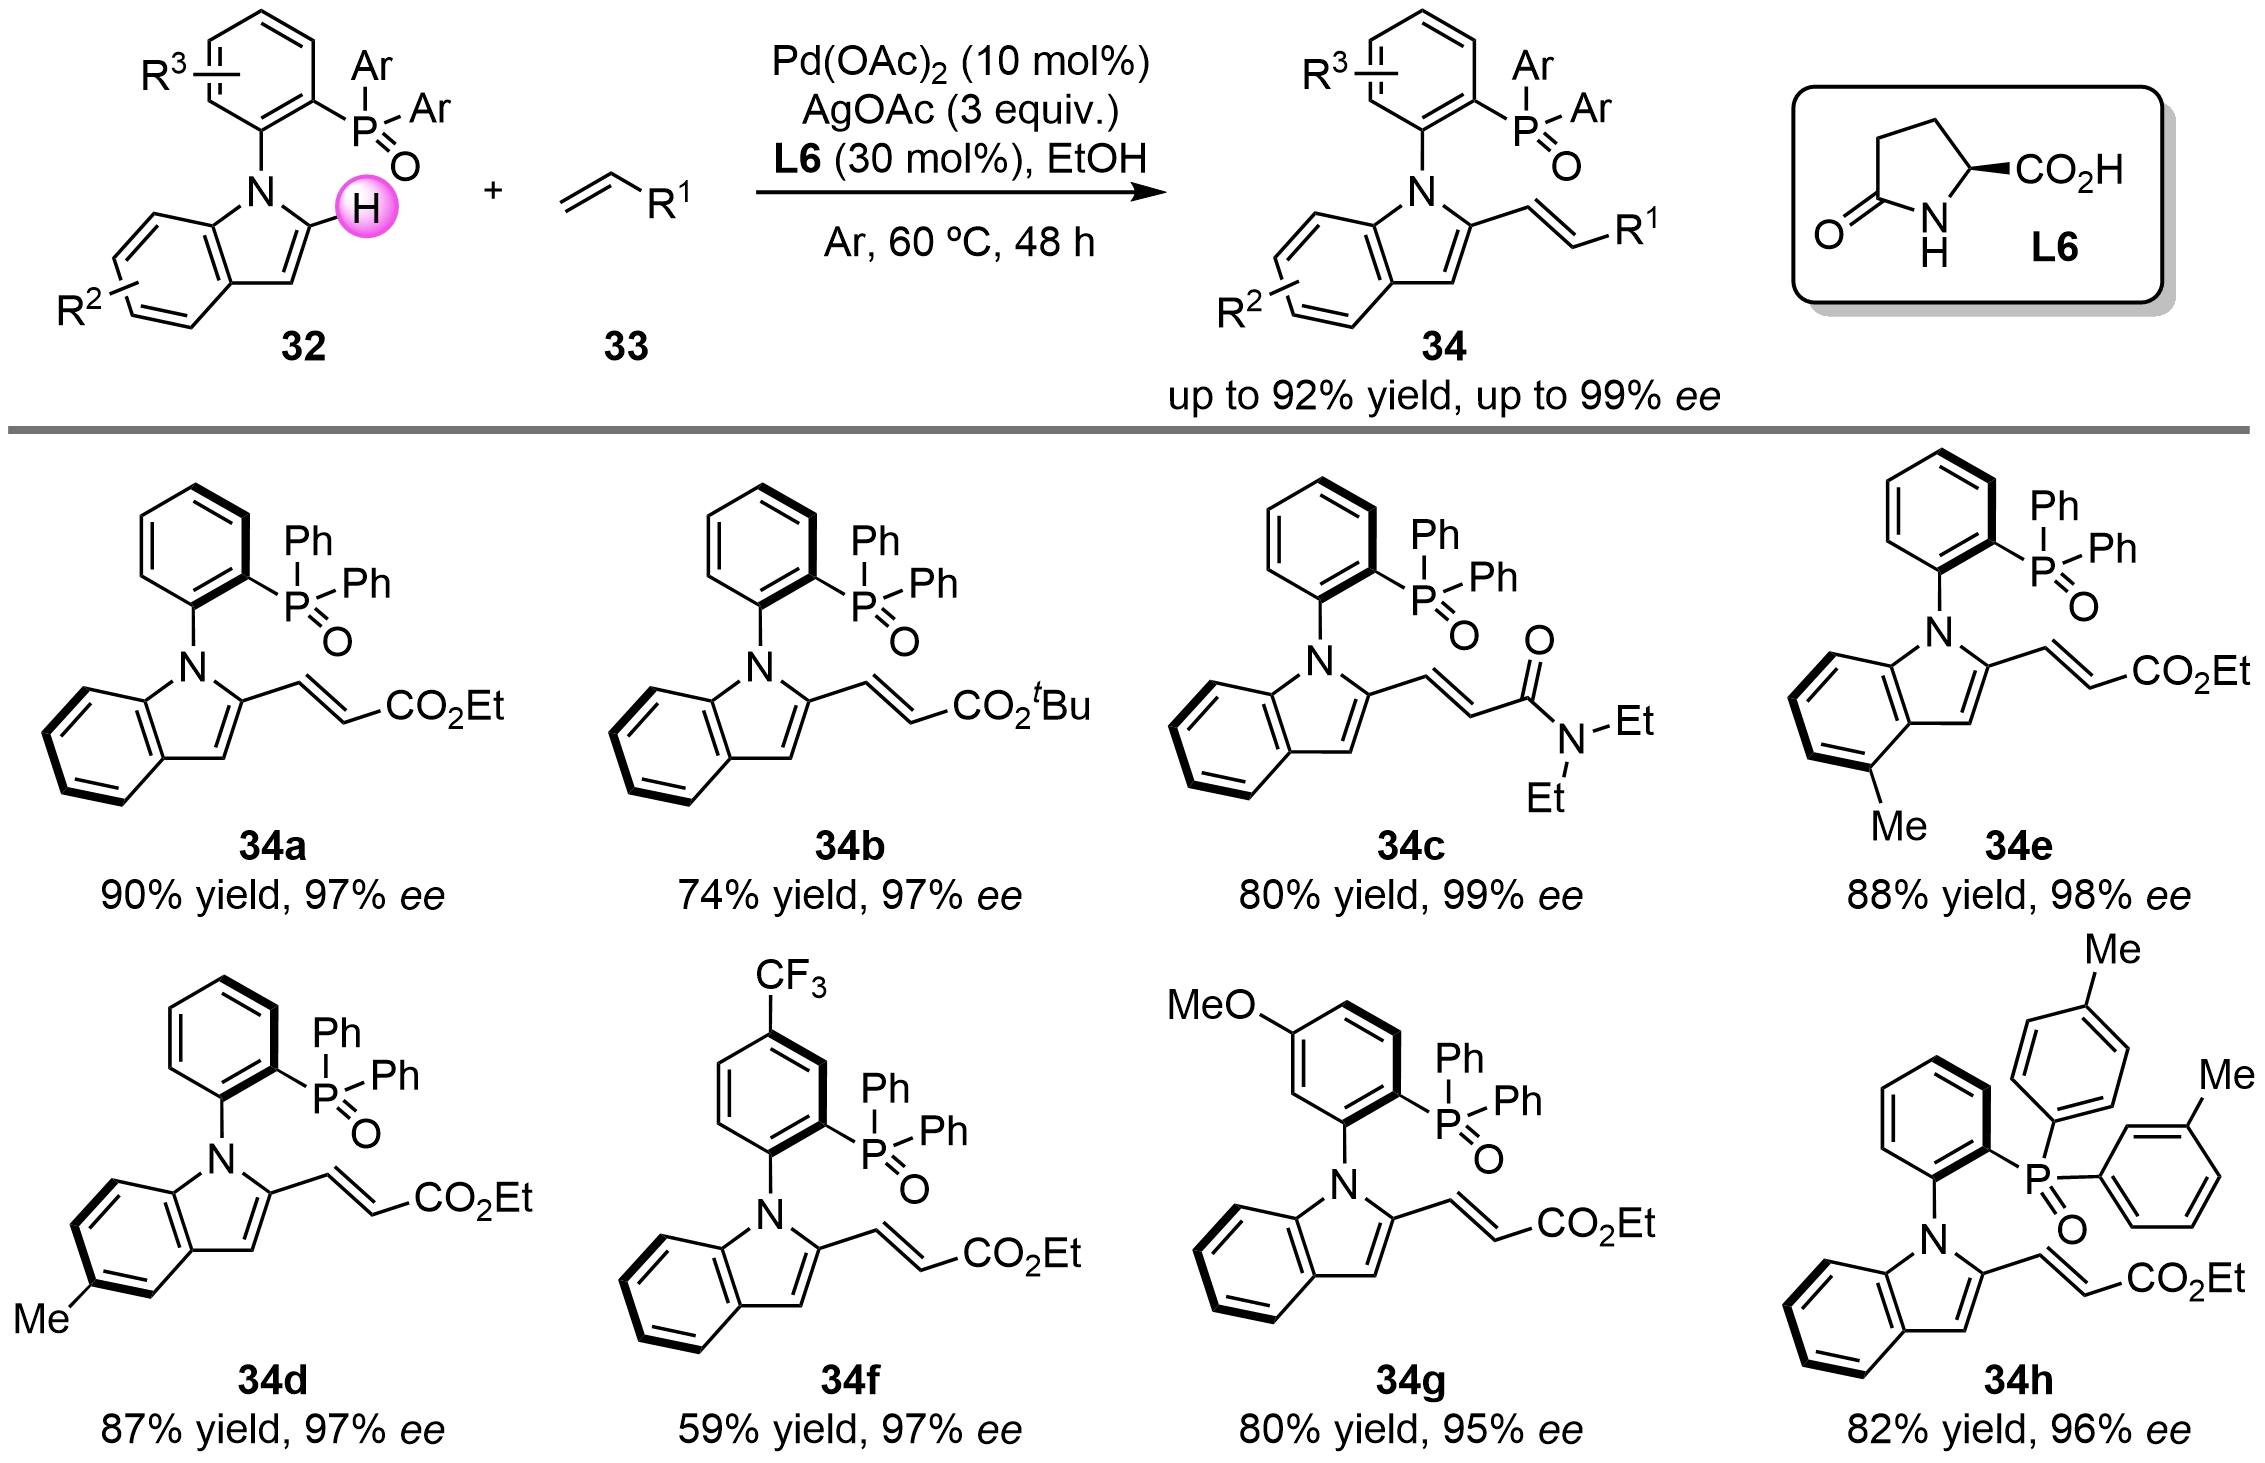
Scheme 14. Construction of C−N axial chirality by palladium-catalyzed asymmetric C−H olefination.
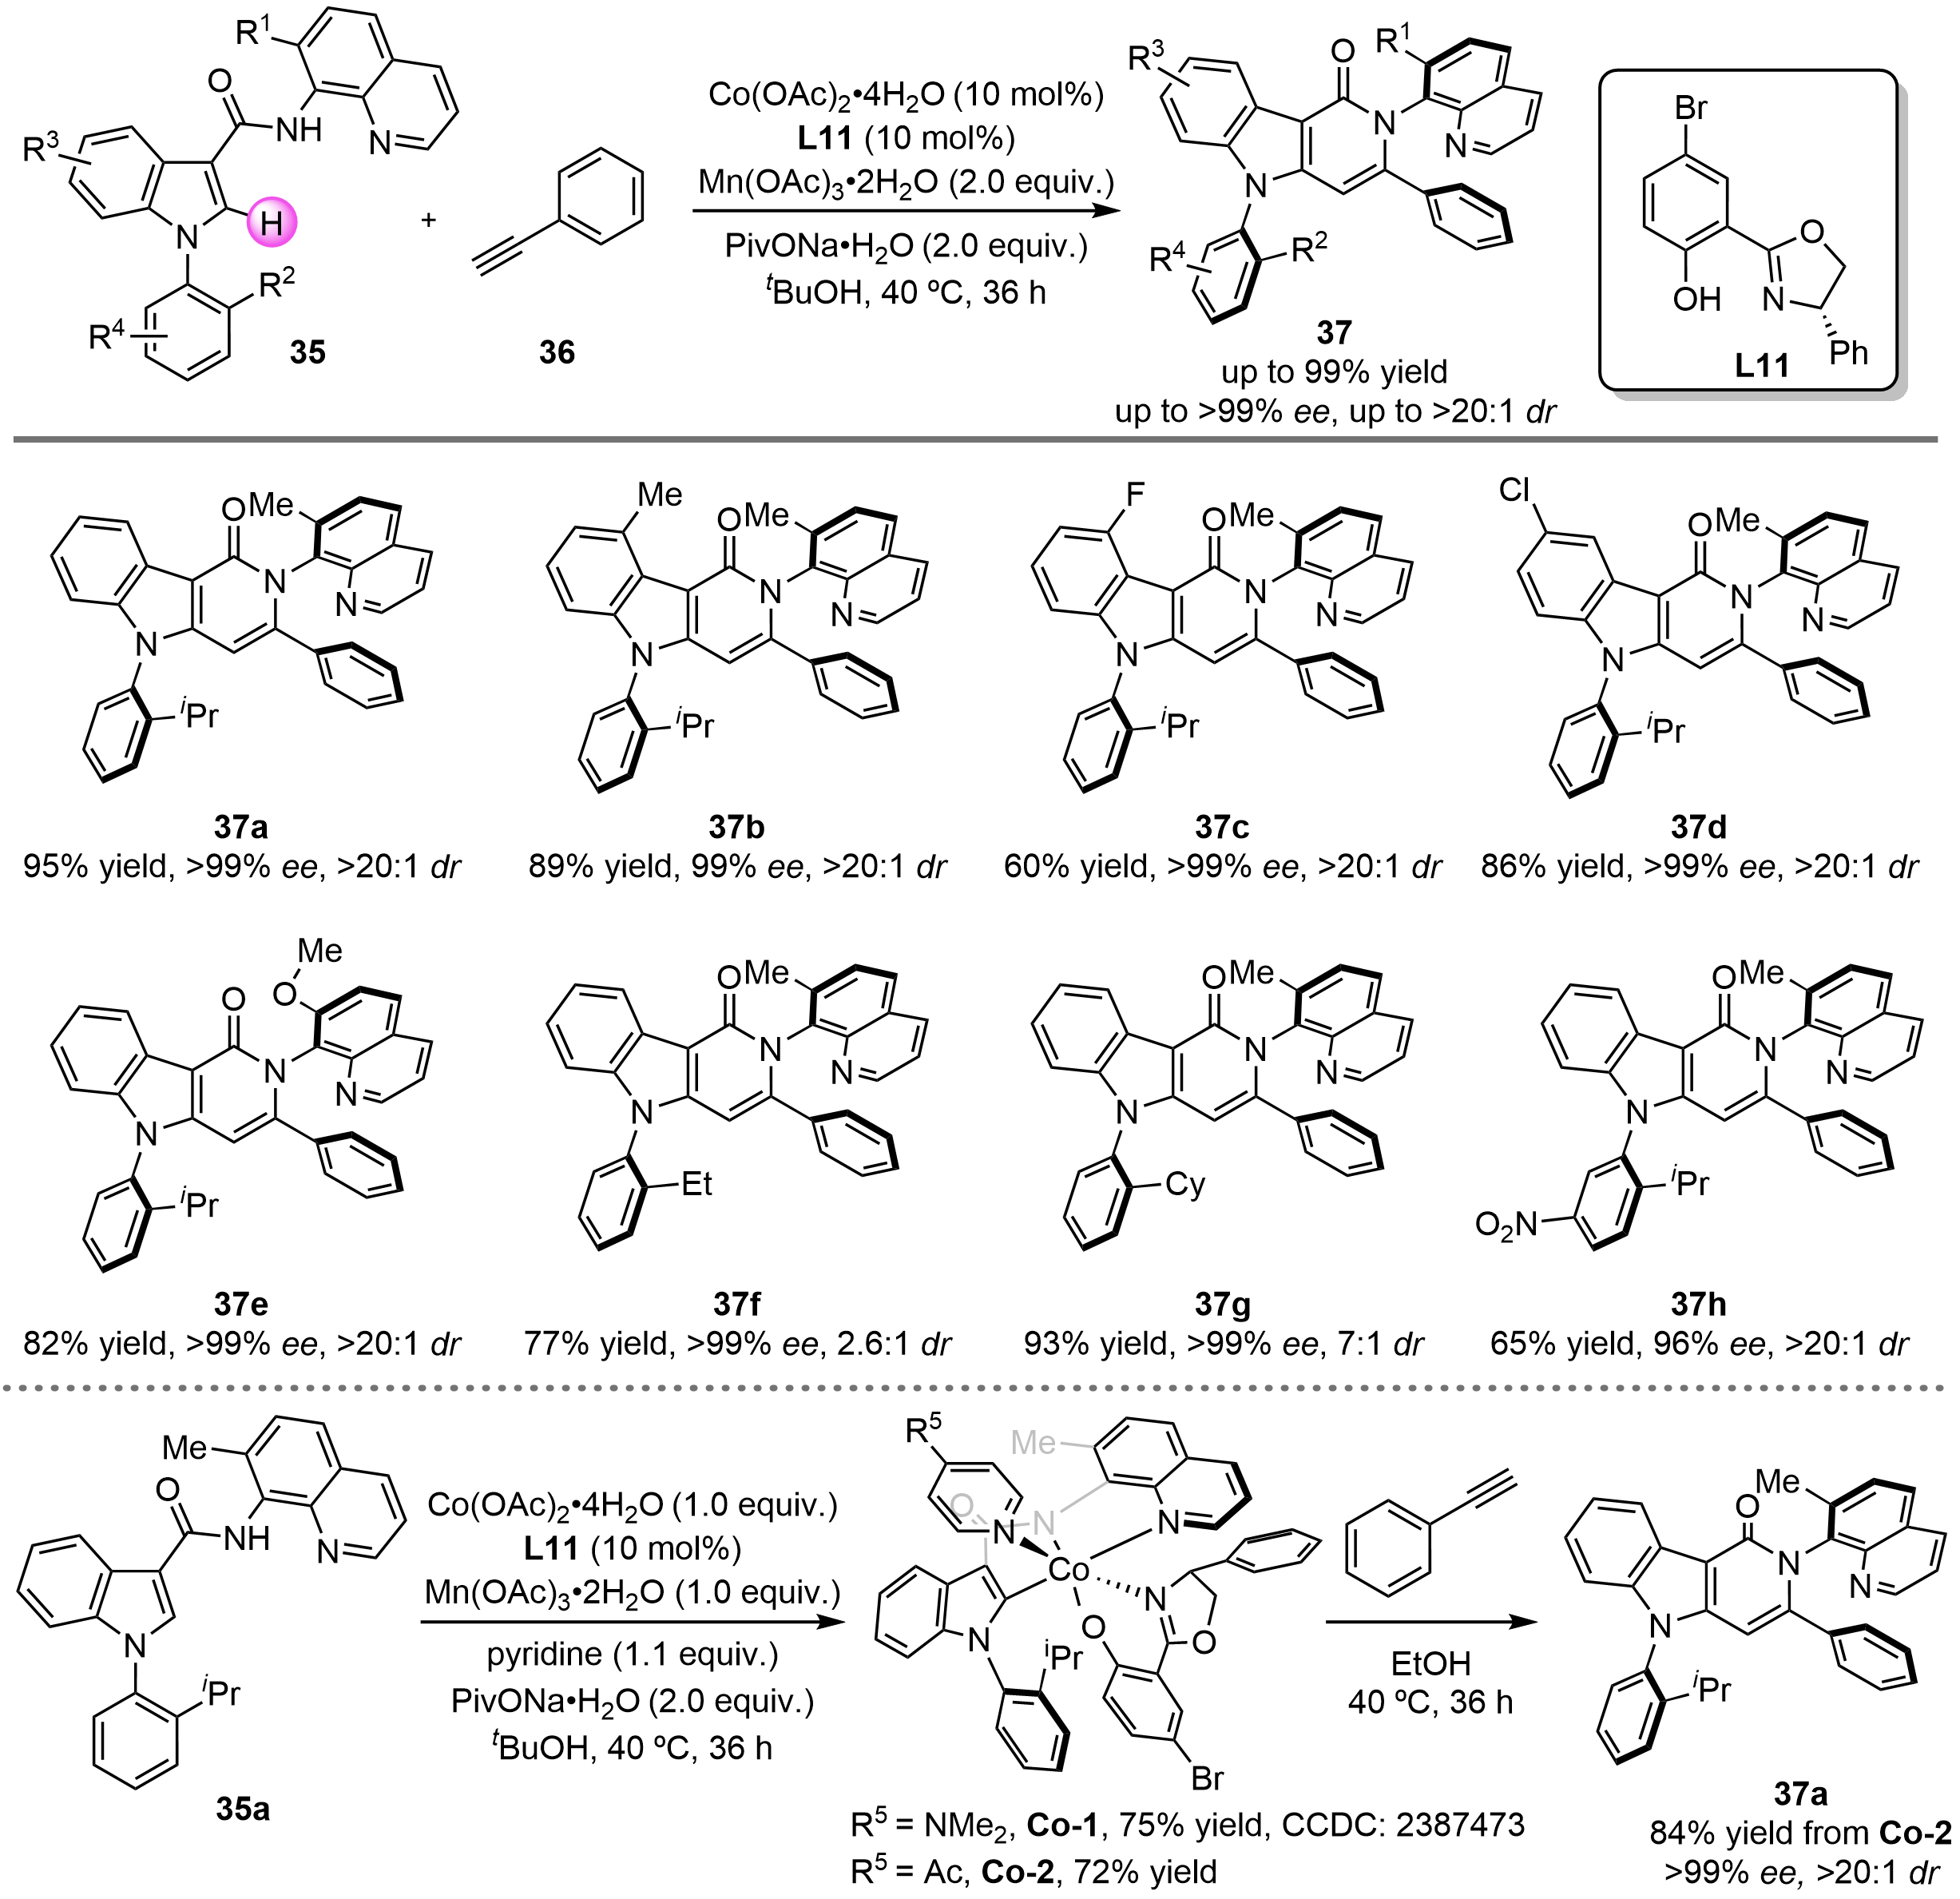
In 2025, the Shi group developed a cobalt-catalyzed system for enantioselective C−H activation. Herein, they present an atroposelective synthesis of chiral pyridoindolones, wherein an innovative compound containing two chiral C−N axes was constructed with excellent stereocontrol via a two-fold C−H [4 + 2] annulation, featuring both six-five and six-six remote C−N axes generated synchronously (Scheme 15)[63]. In mechanistic studies, pyridine-stabilized cobaltacycles were synthesized and fully characterized. X-ray diffraction revealed that both C−N axes were concomitantly generated and sterically locked during the C−H cyclometalation process. The atropocontrol originates from two π-π stacking interactions, which are the quinolyl-phenyl and N-aryl-salicyl interactions. These interactions are crucial for stabilizing the chiral axes.
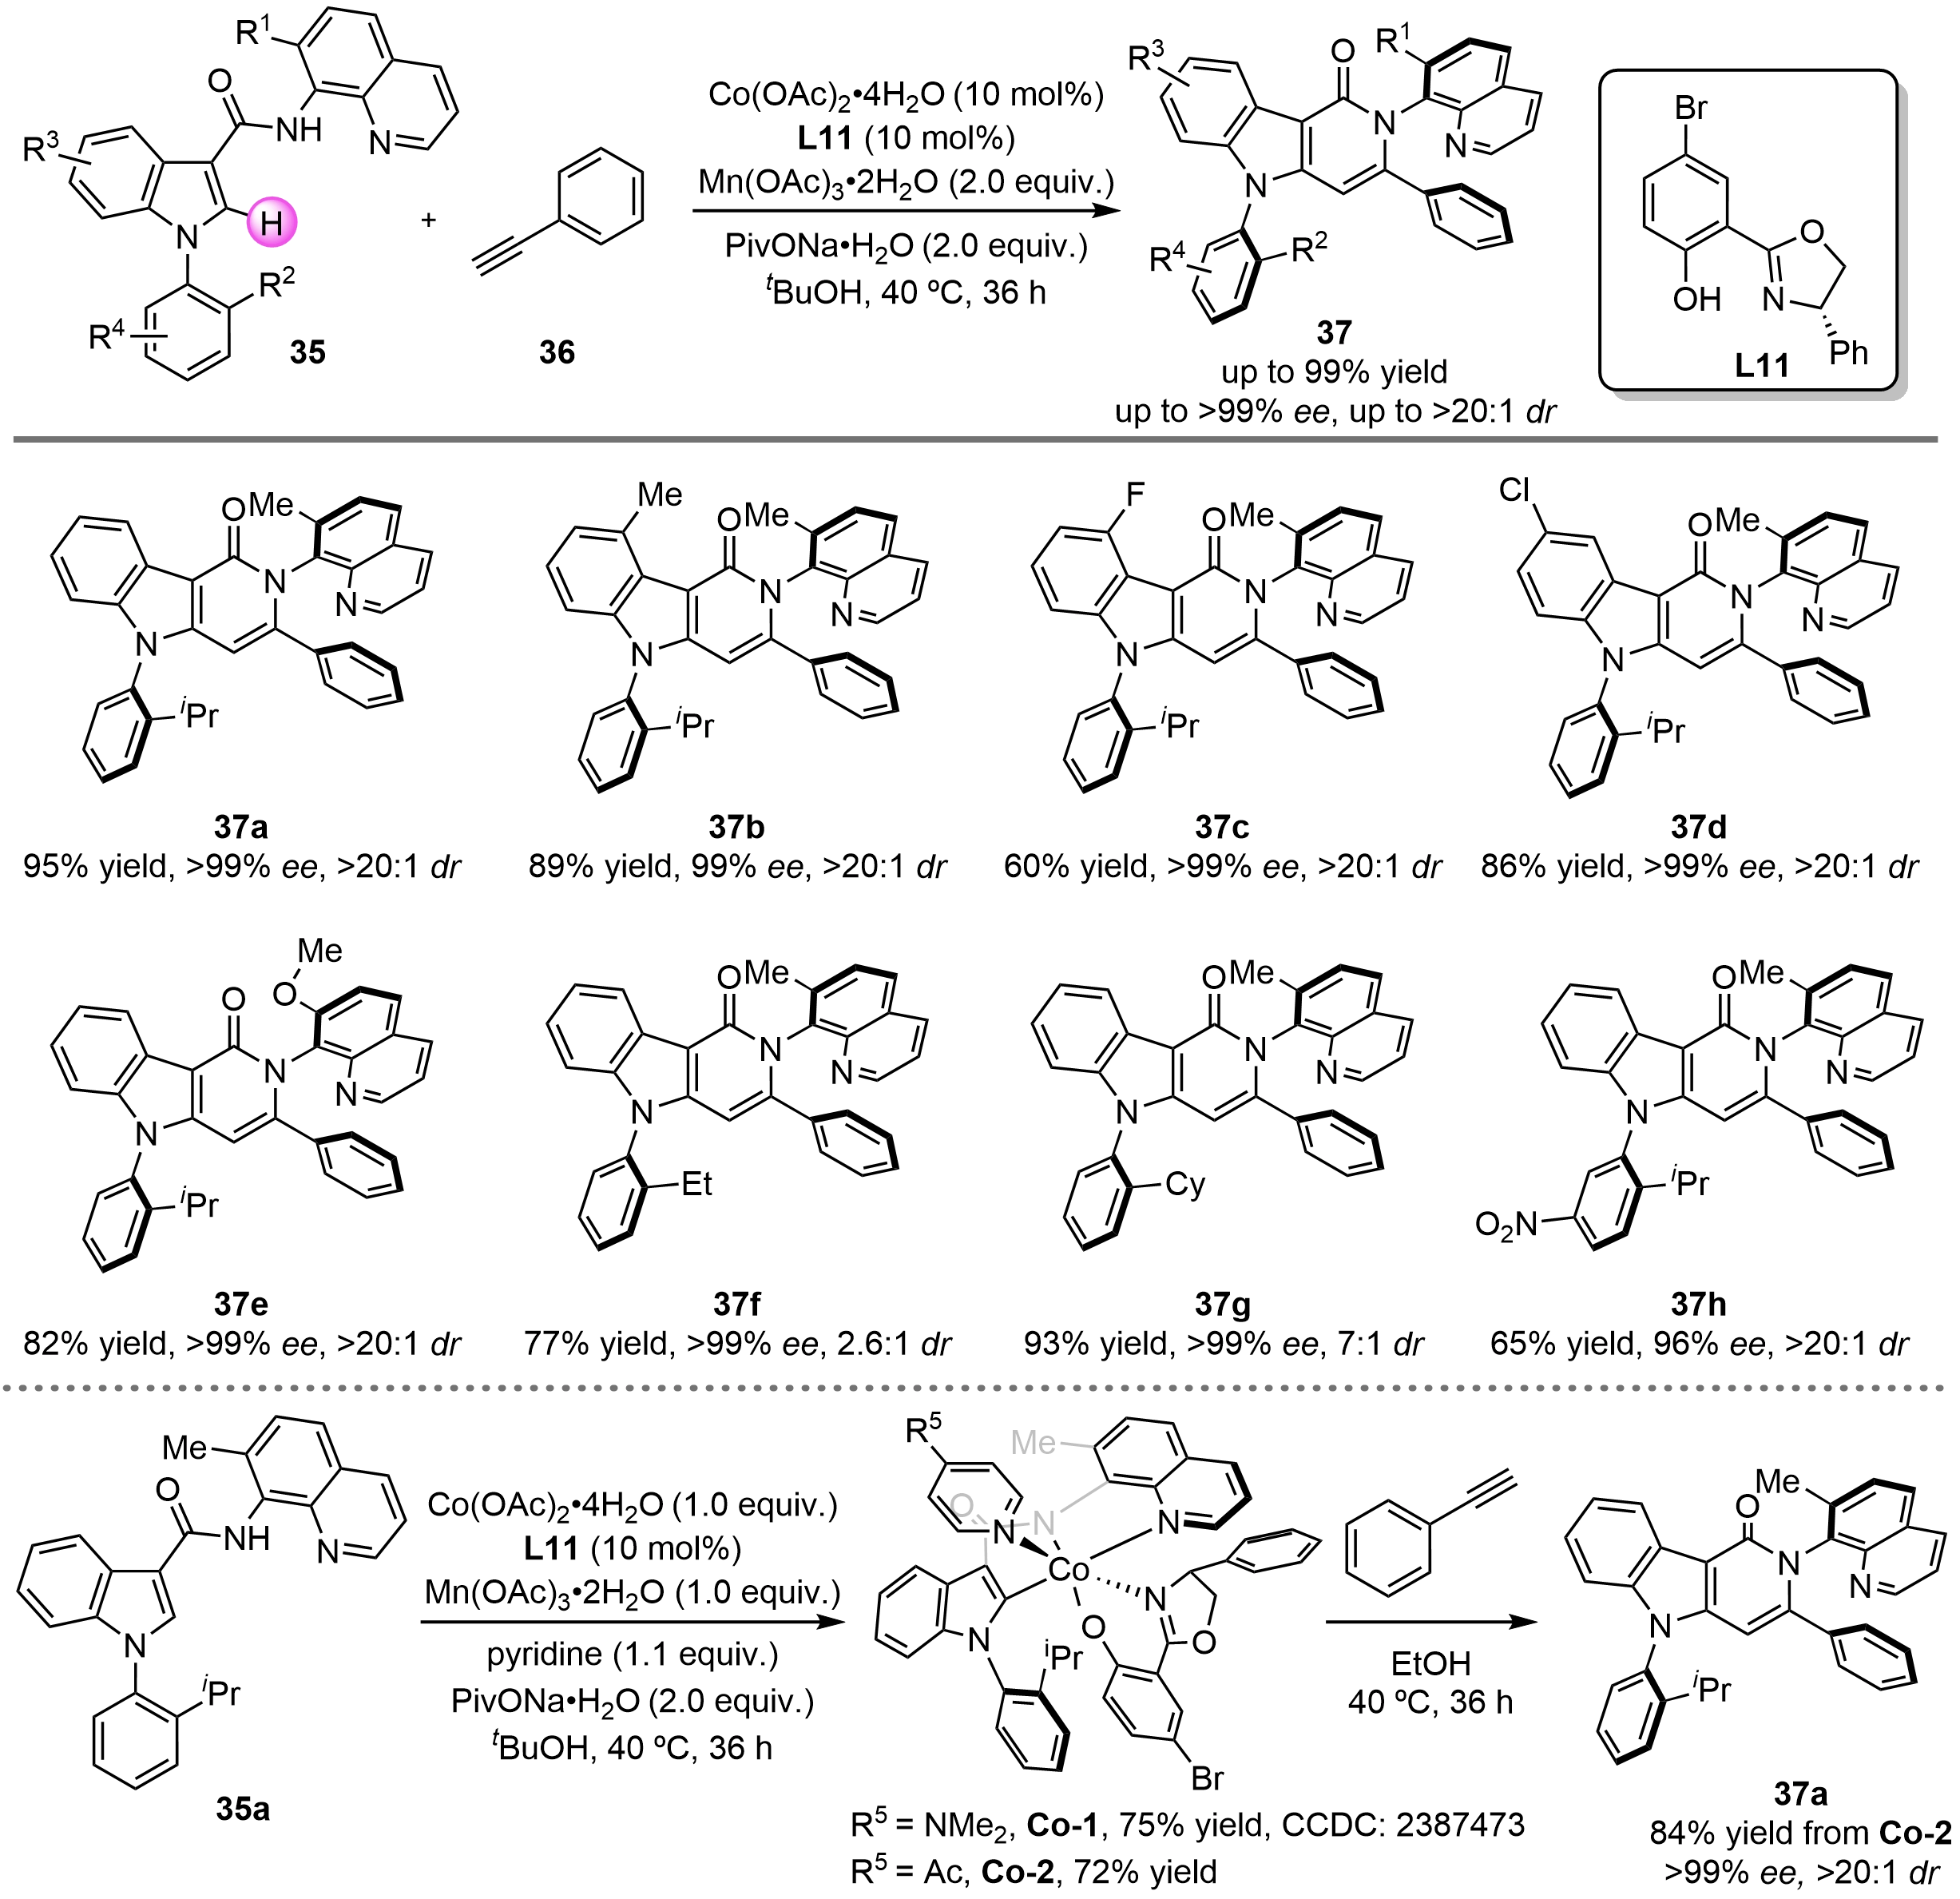
Scheme 15. Construction of C−N axial chirality by cobalt-catalyzed asymmetric C−H activation.
2.2 DKR of biaryls with free axial rotation via C−H functionalization with a transient directing group
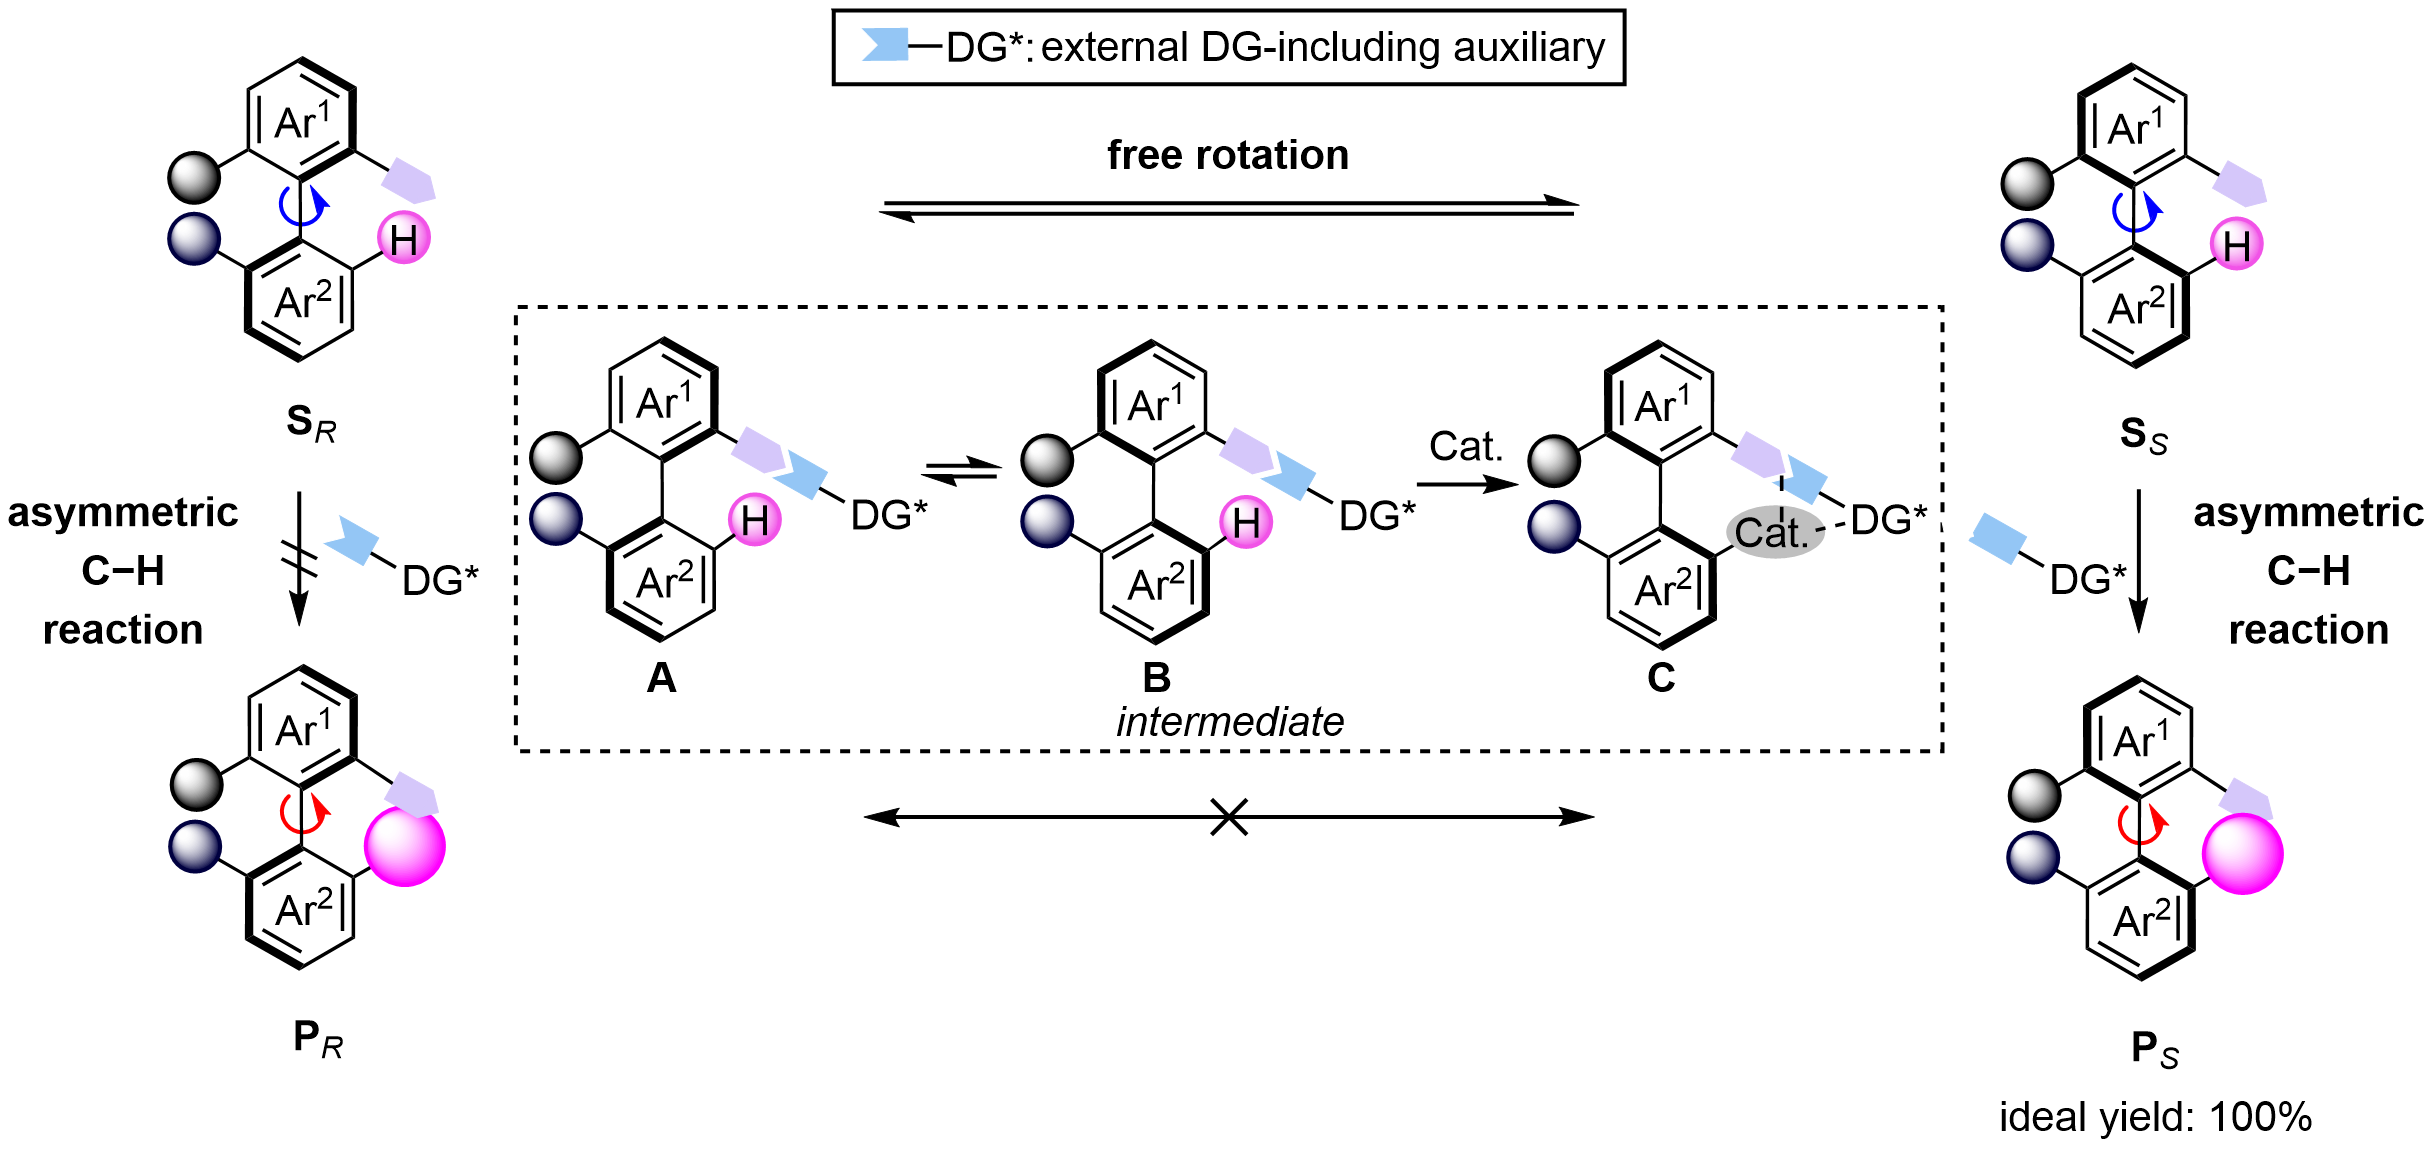
In the aforementioned examples, it is evident that the DKR of biaryls through the direct C−H functionalization strategy has been effectively developed to accomplish the enantioselective synthesis of axially chiral biaryl compounds. However, this approach often necessitates the pre-installation of a suitable directing group on the substrate structure. The directing group is not an indispensable moiety for the construction of biaryl skeletons, which generally has to be removed afterwards. In contrast, the strategy utilizing a transient directing group allows for a reversible connection between an external DG-including auxiliary and the substrate, thus avoiding the prior installation of a directing group. The Shi group investigated biaryl substrates bearing an ortho-formyl group, which can reversibly react with an external amino acid to transiently install an imine moiety onto the substrate framework (A and B). Crucially, the ingenious utilization of an amino acid as a co-catalyst introduces an additional carboxylic acid group, enabling the transient imine and carboxylate to cooperatively function as a bidentate directing group. Subsequently, the preferred diastereomer B selectively coordinates with the transition metal to undergo C−H cleavage, thus affording an axially stereoenriched biaryl palladacycle intermediate C. This intermediate promptly undergoes subsequent asymmetric transformation to deliver the desired axially chiral biaryls with a theoretical yield of up to 100% (Scheme 16).
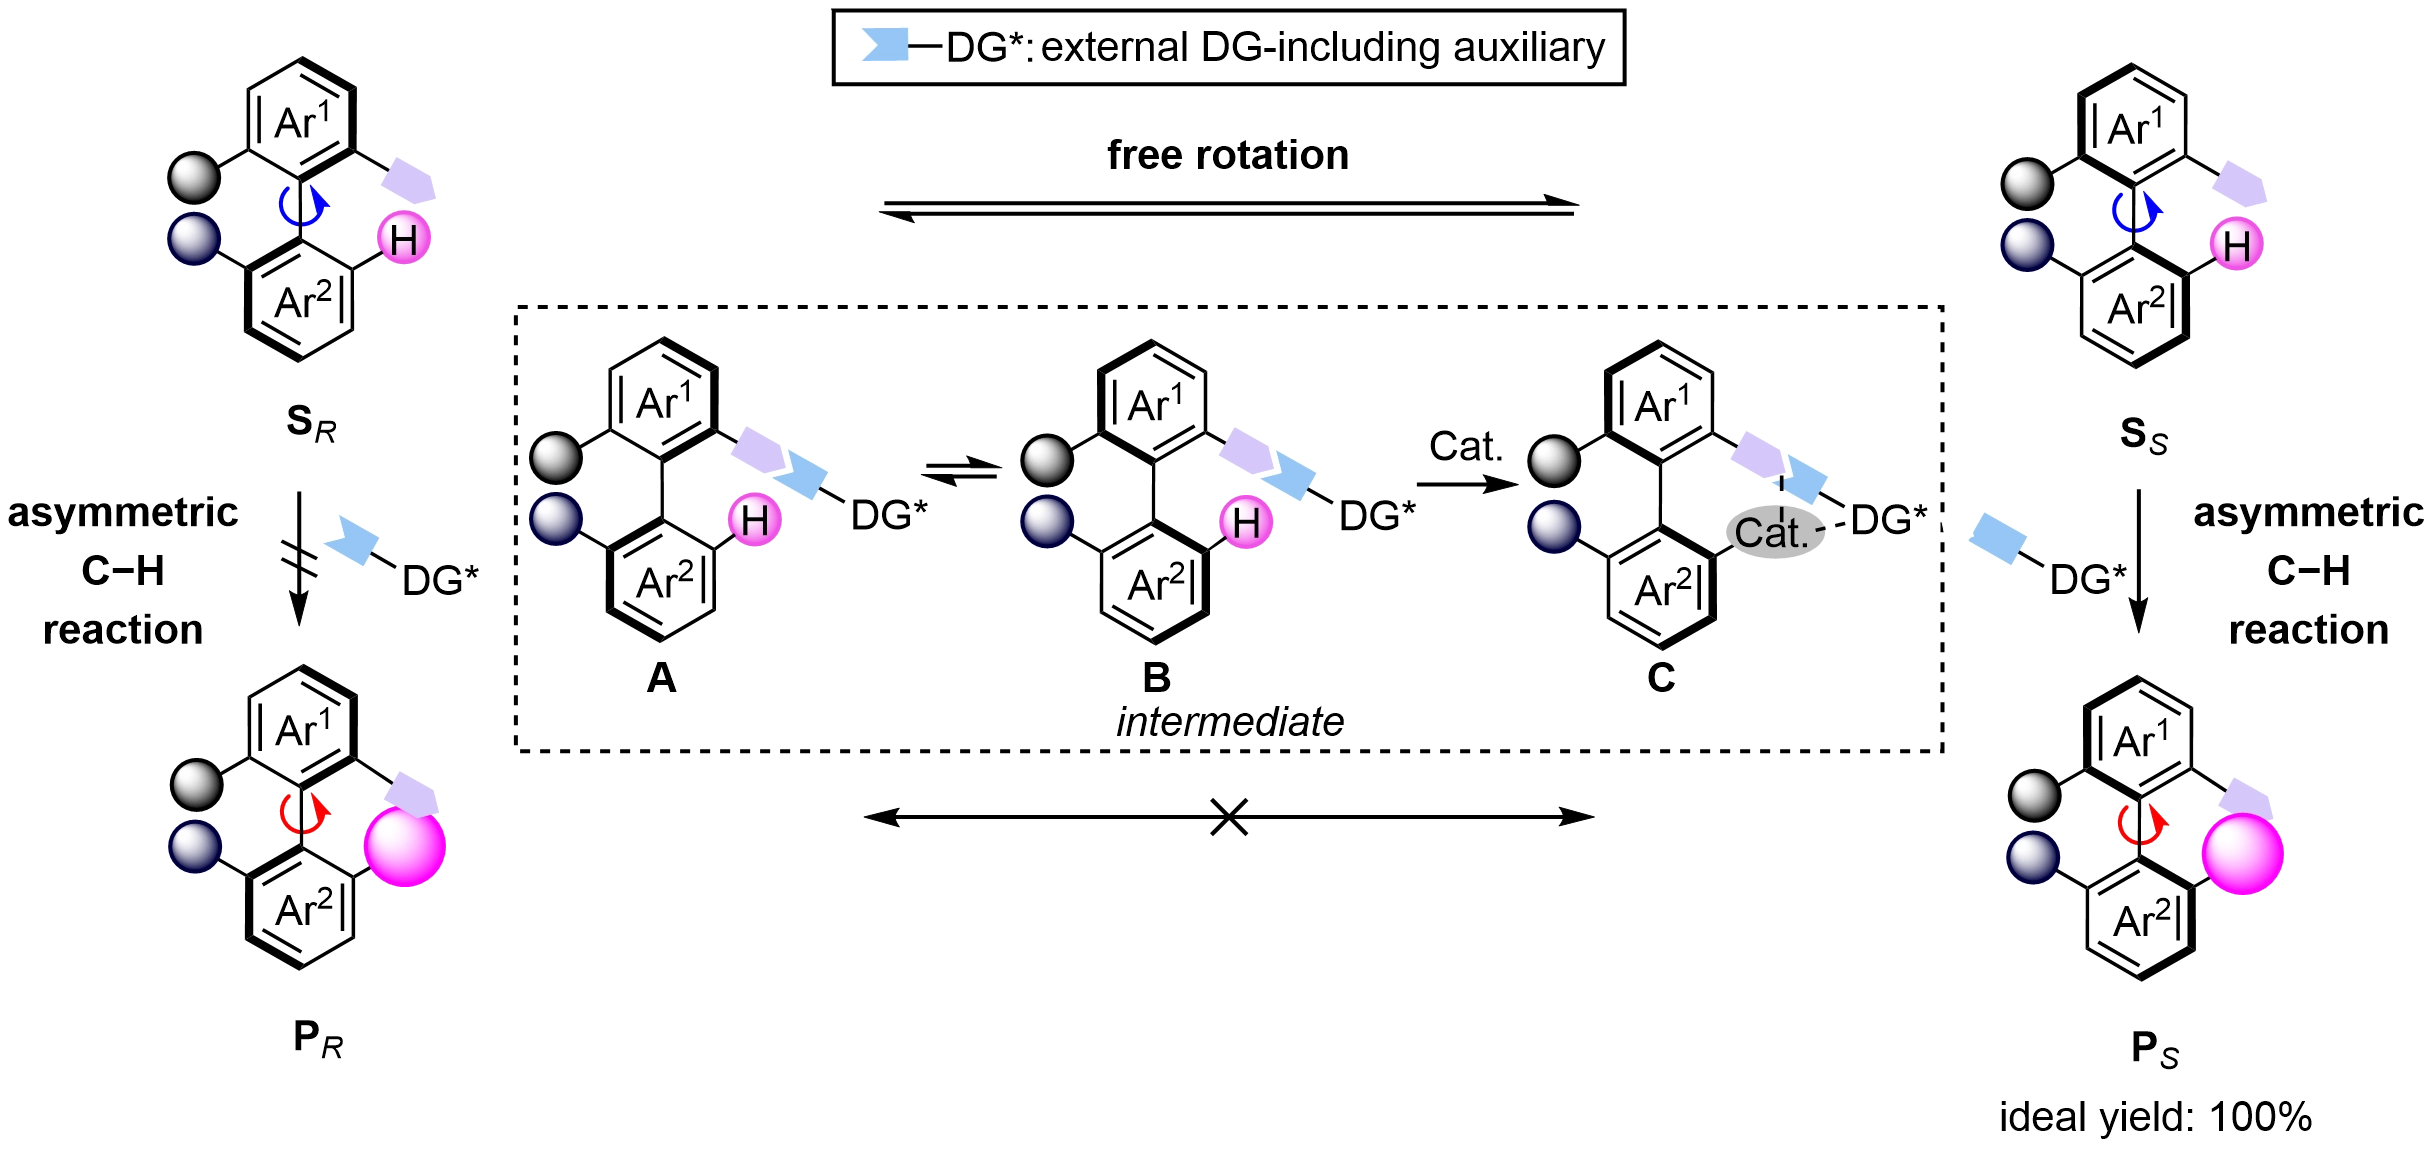
Scheme 16. DKR of biaryls via C−H functionalization by using an external transient directing group. DKR: dynamic kinetic resolution.
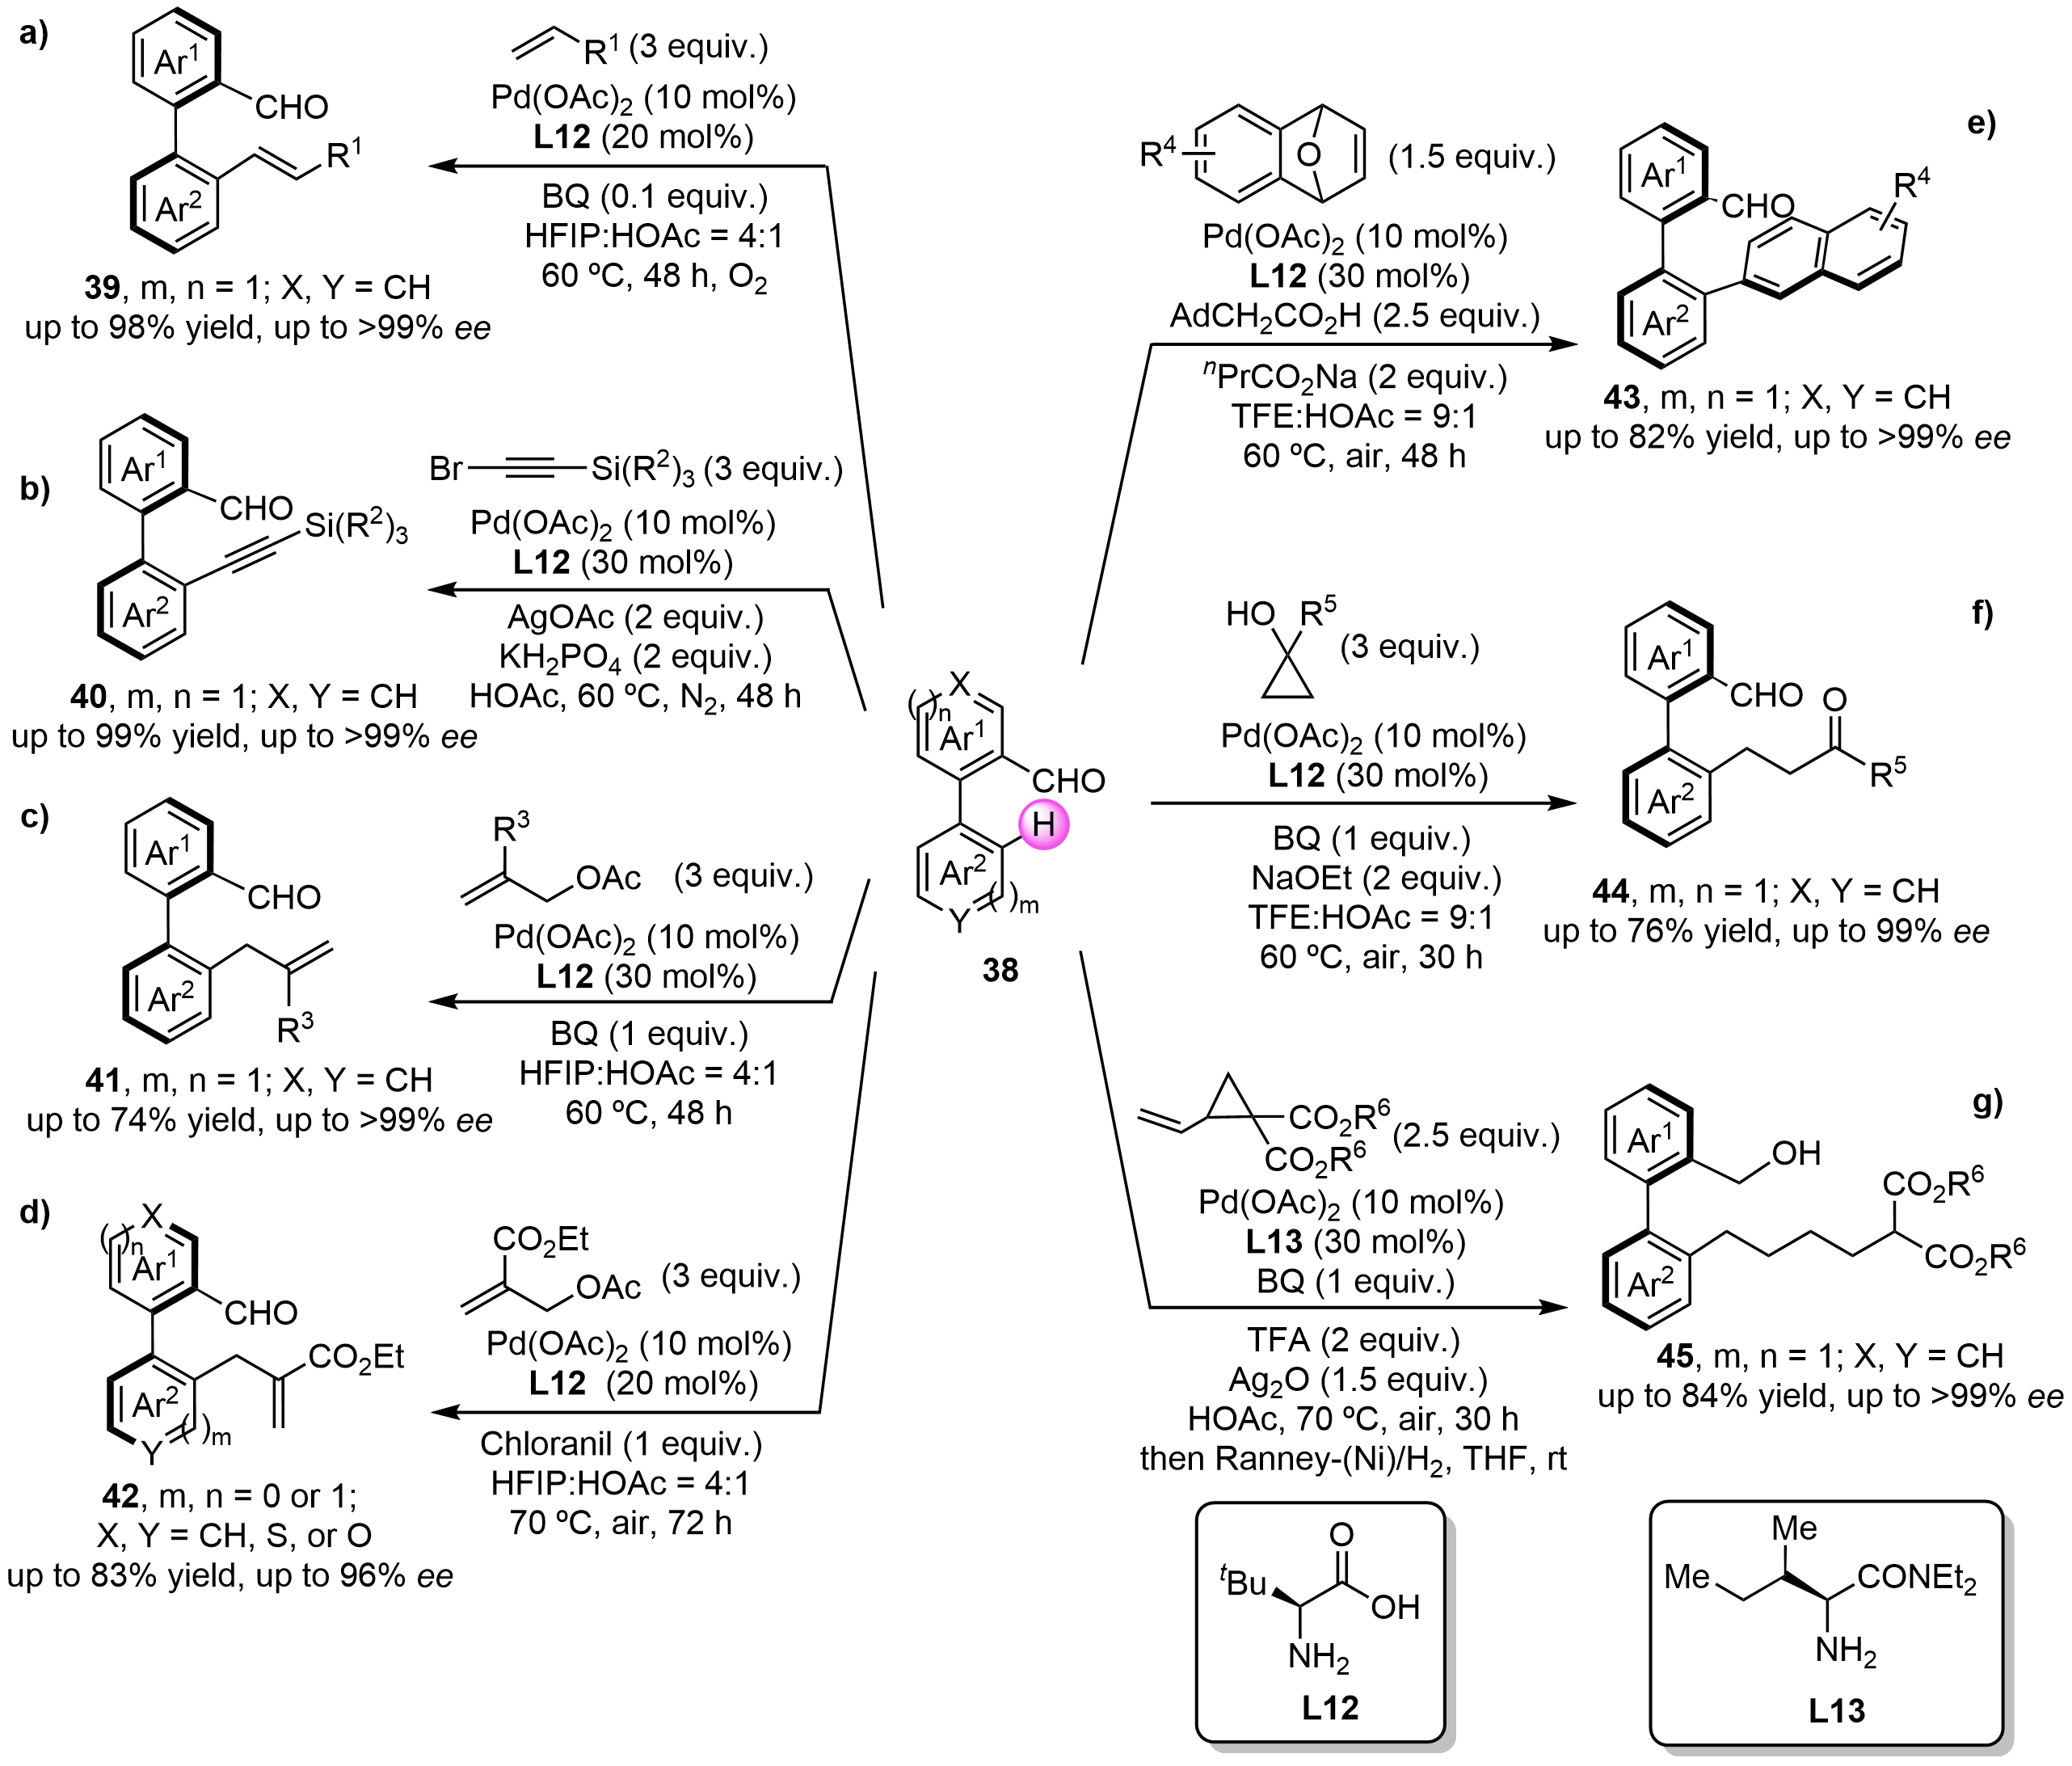
From 2017 to 2021, the Shi group made significant progress in addressing the asymmetric construction of biaryl axial chirality via C−H functionalization by using a transient directing group. For instance, the palladium-catalyzed oxidative Heck reaction with terminal olefins afforded products 39 in high yield with excellent enantioselectivity (Scheme 17a)[64]. The coupling reaction with alkynyl bromides under redox-neutral reaction conditions led to enantioenriched products 40 efficiently (Scheme 17b)[65]. It is worth noting that the asymmetric alkynylation method was successfully applied in the stereocontrolled synthesis of (+)-isoschizandrin and (+)-steganone on a gram-scale. Later, they demonstrated that the C−H allylation reaction with allyl acetate could be achieved via β-oxygen elimination to form allylated chiral biaryls 41 and 42, respectively (Scheme 17c,d)[66,67]. 7-Oxabenzonorbornadienes[68] and cyclopropanes[69] were also adopted for achieving selective C−H naphthylation and alkylation of biaryls by using a similar strategy (Scheme 17e,f,g). In 2025, the Jeganmohan group reported an atroposelective C−H vinylation protocol employing a vinylsilane under palladium-catalyzed C−H activation[70].
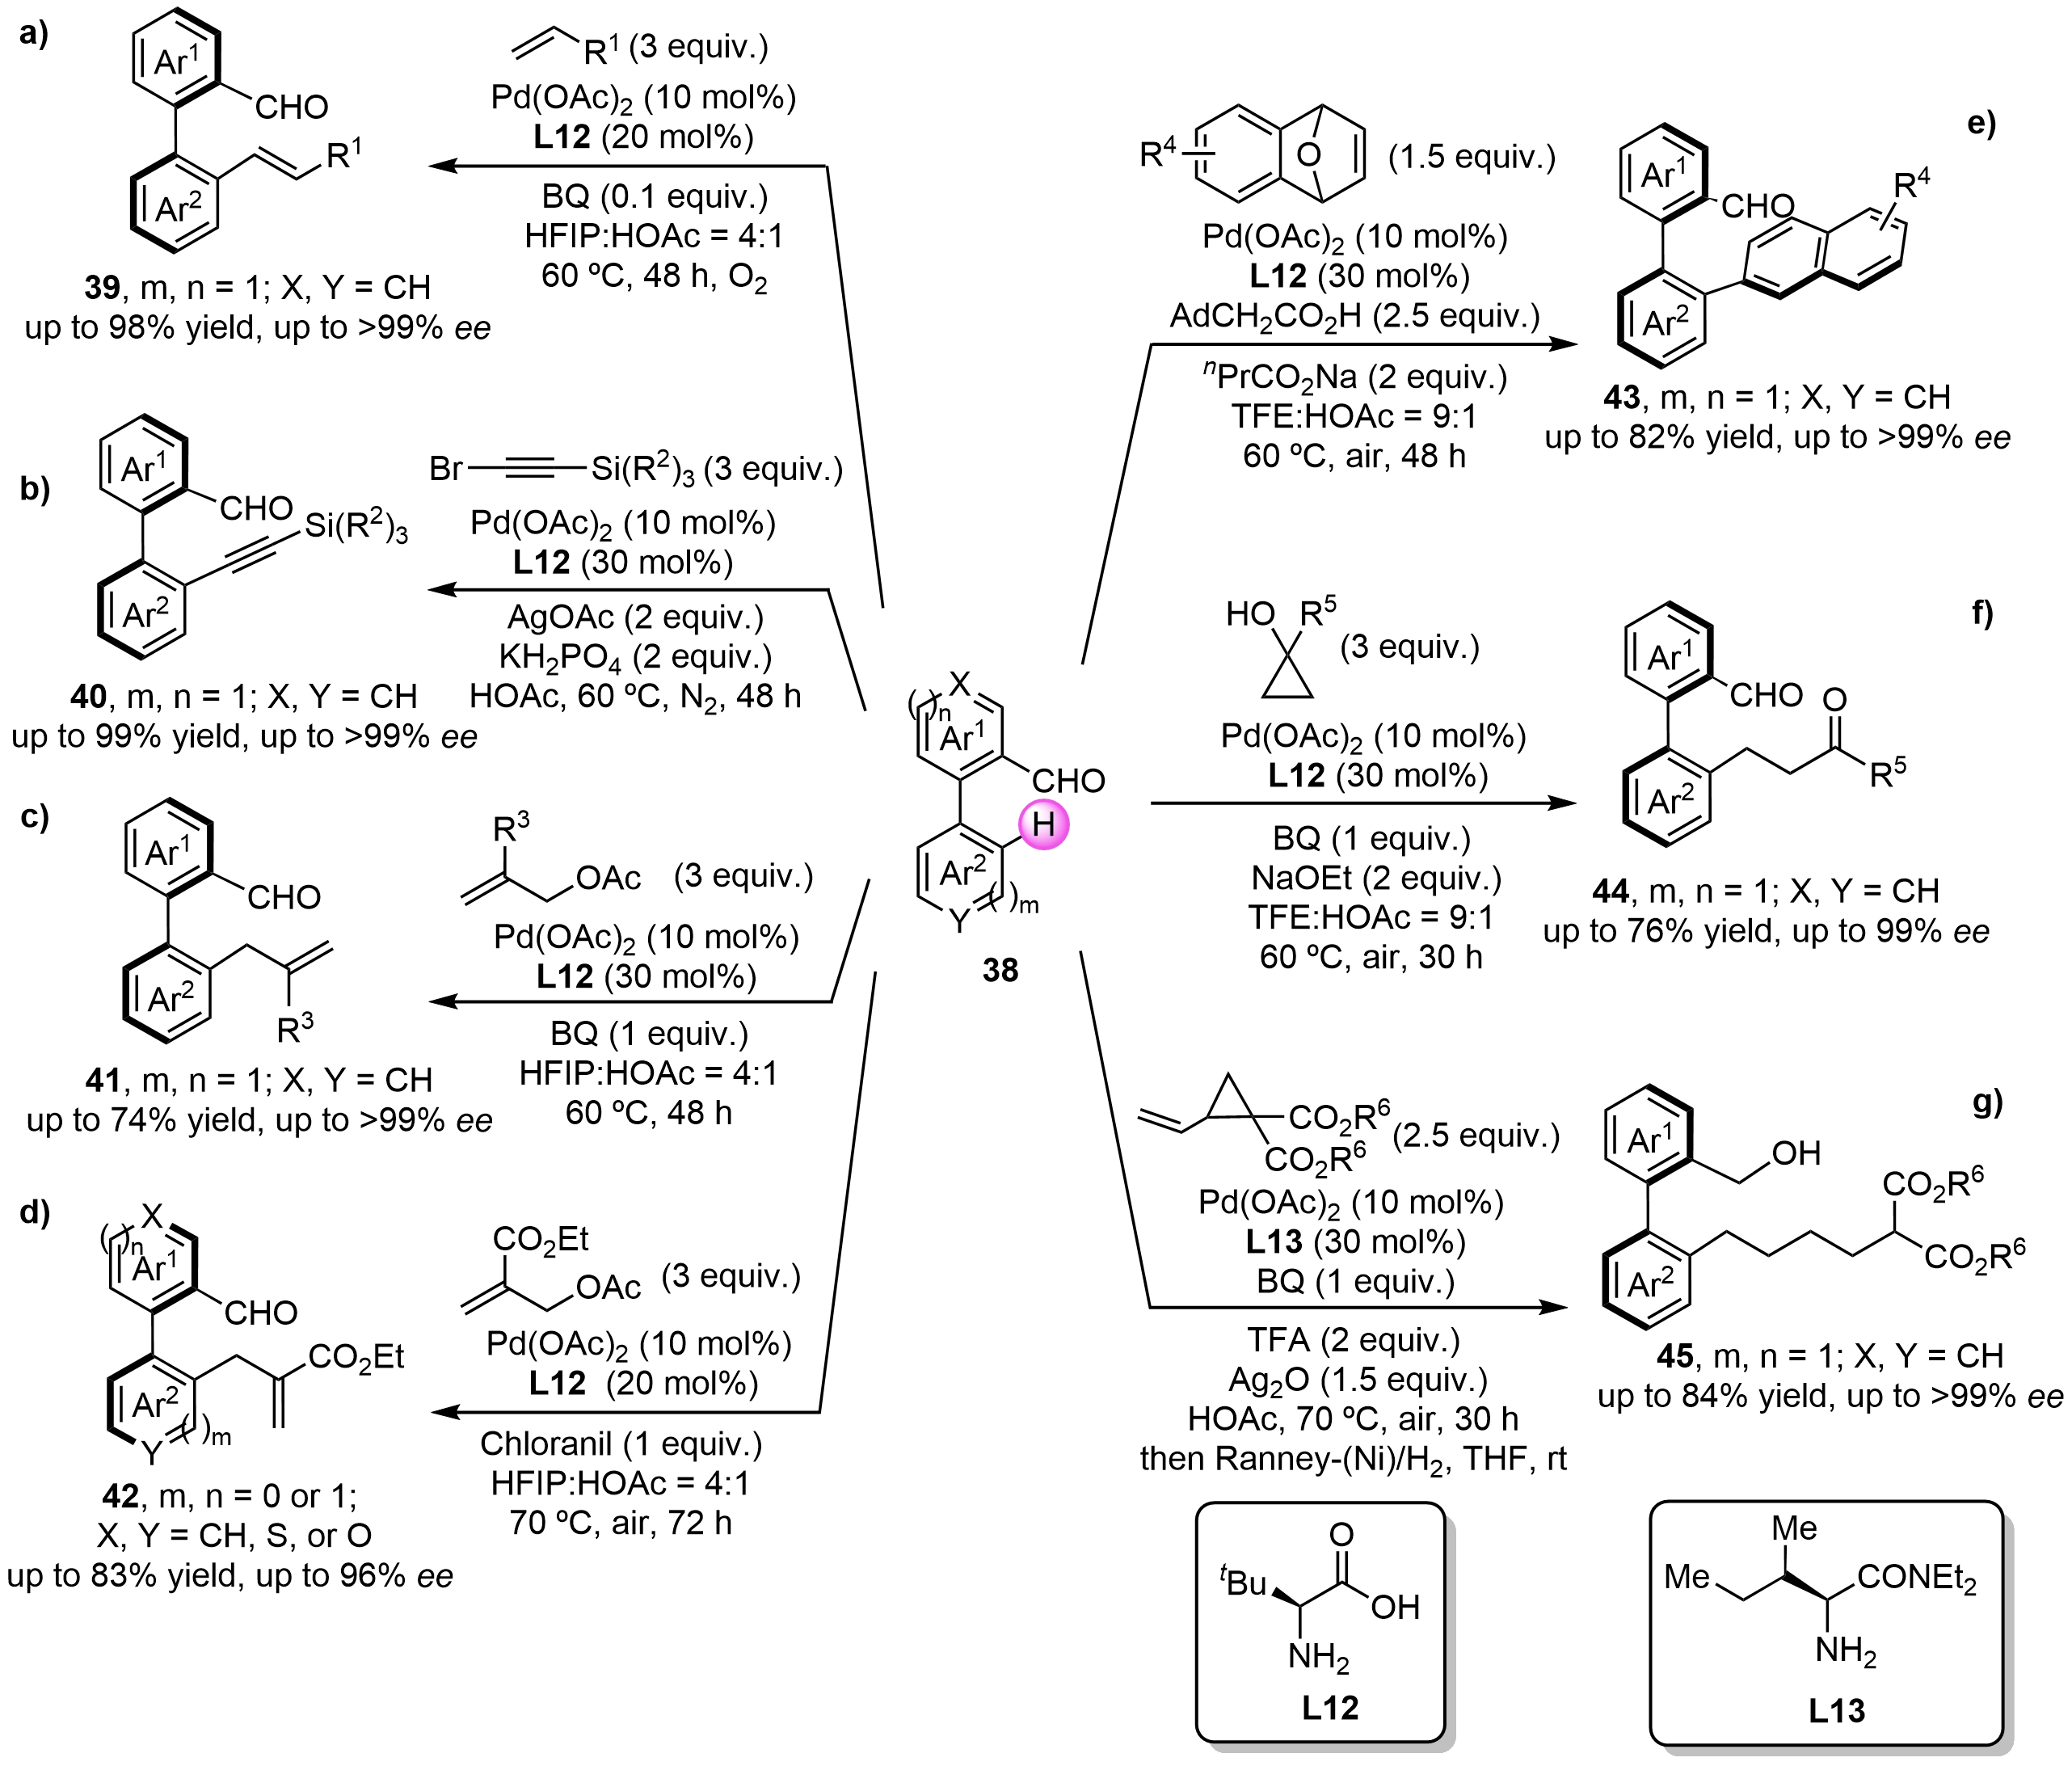
Scheme 17. Construction of C−C axial chirality by palladium-catalyzed asymmetric C−H functionalization with a transient chiral auxiliary.
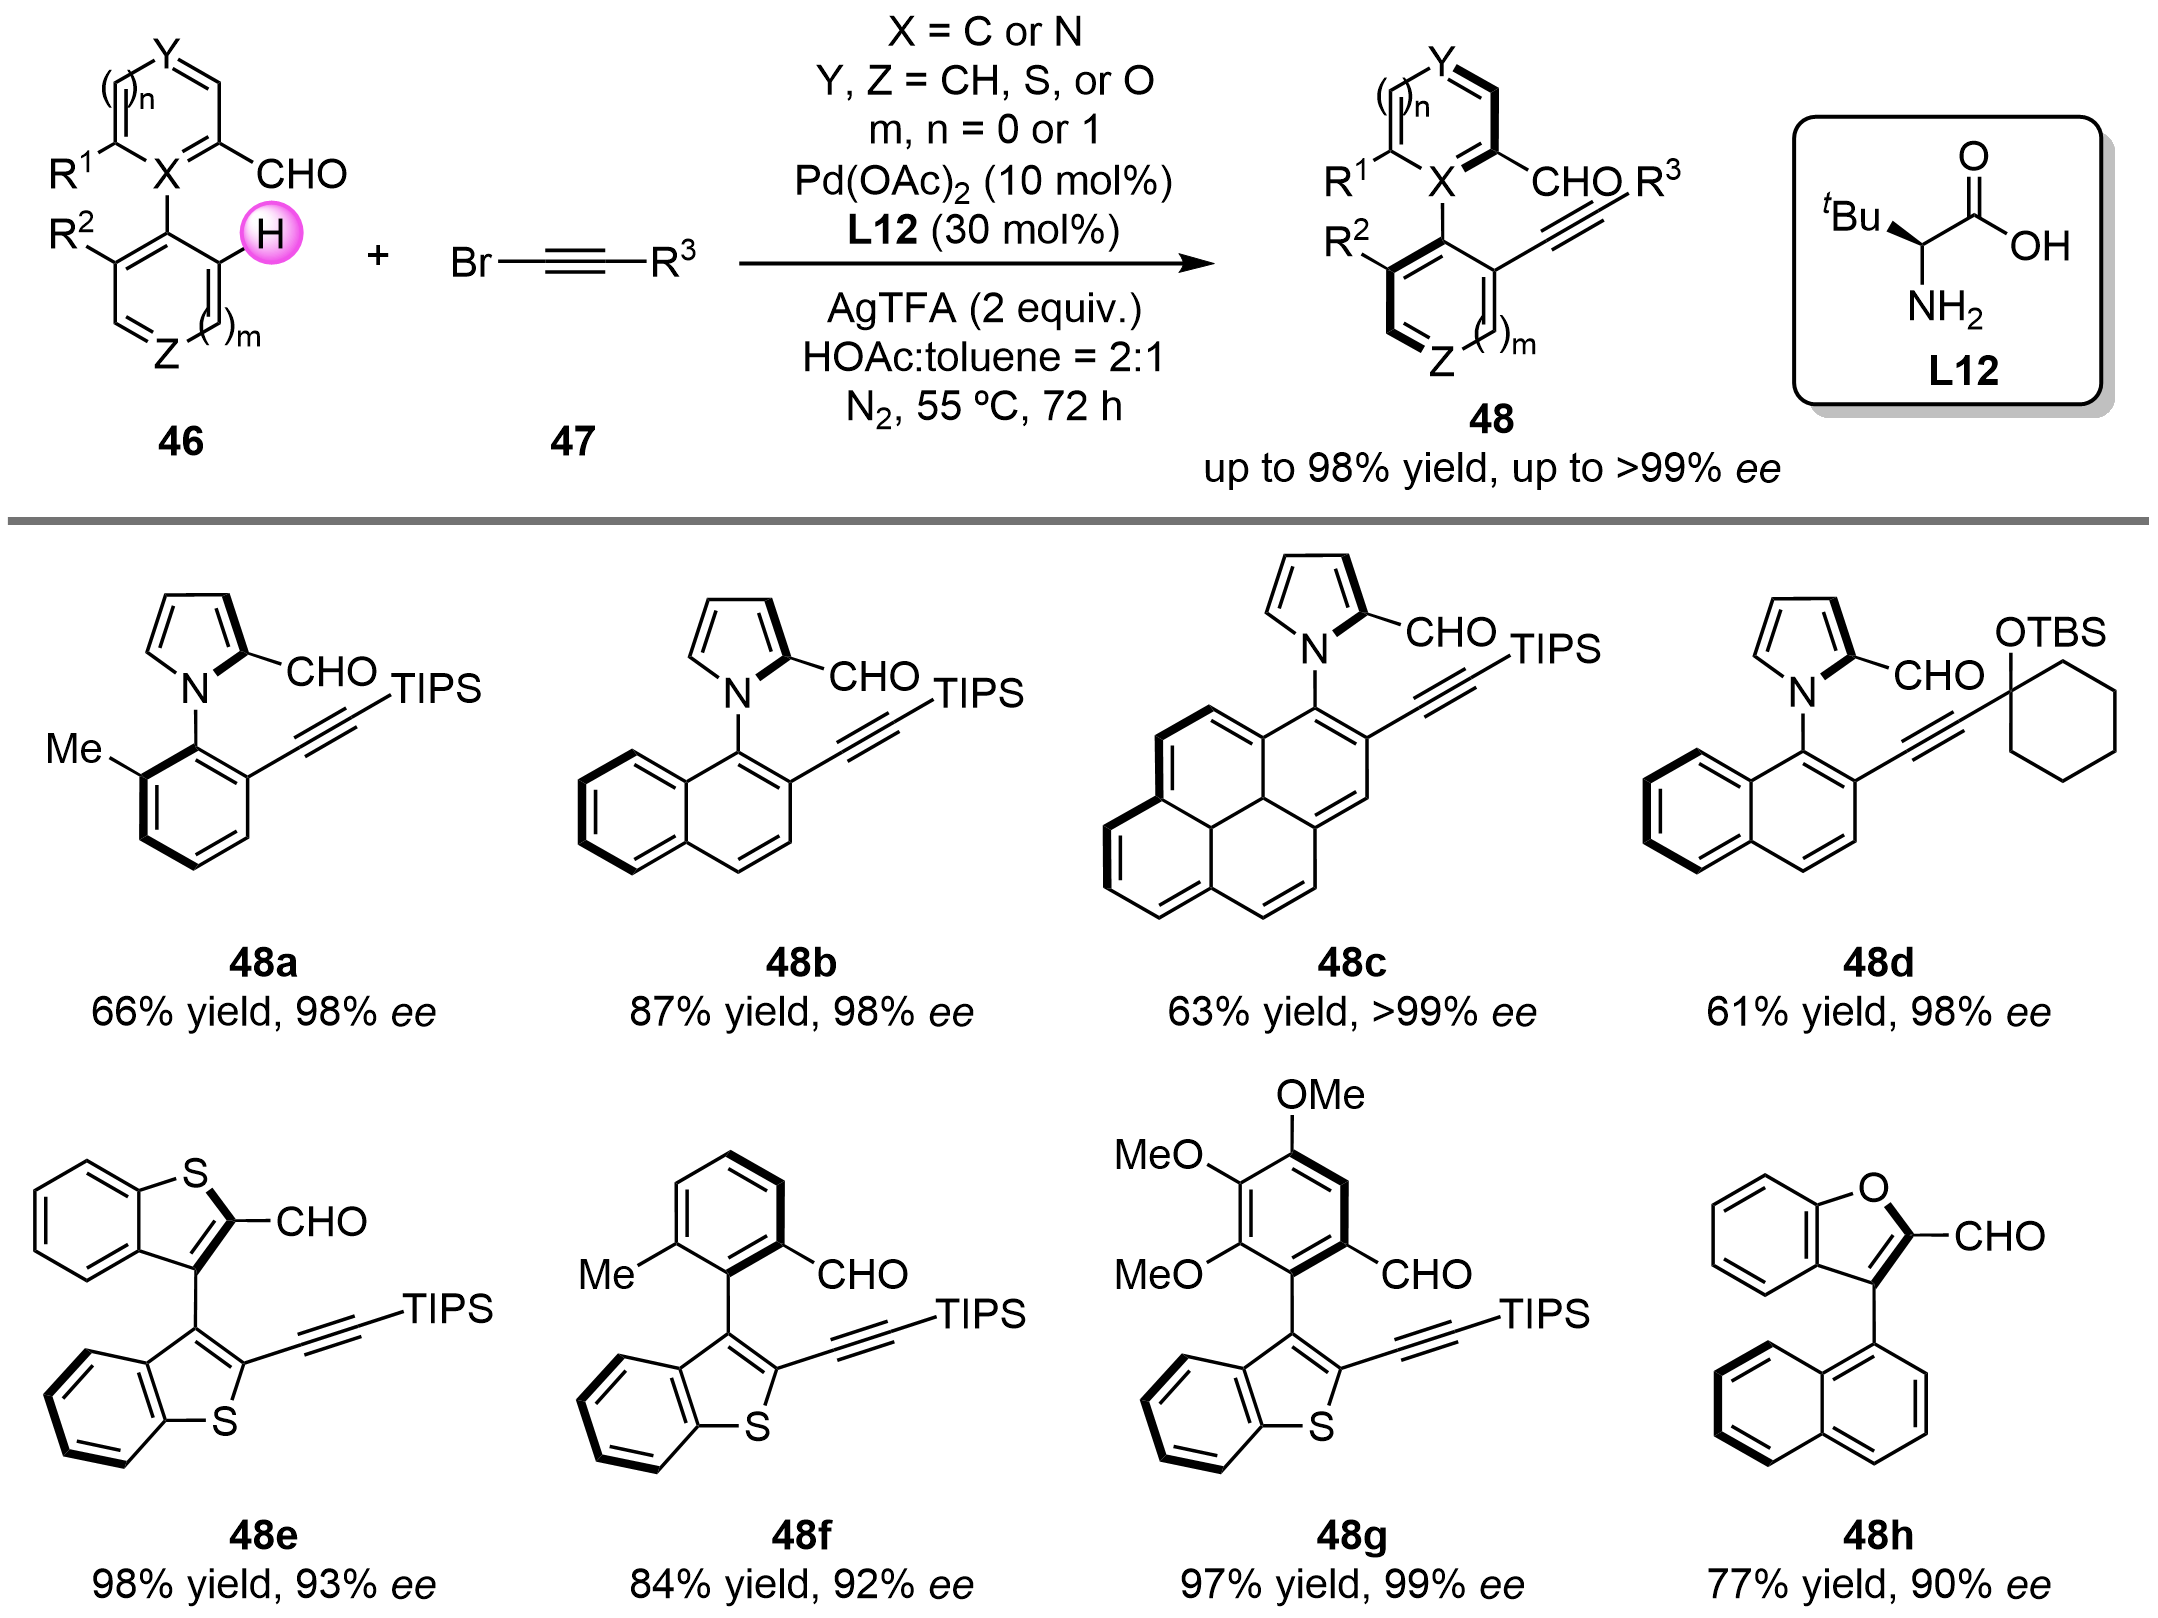
In addition to the construction of C−C axial chirality, the Shi group also focused on the synthesis of C−N axial chirality, and they successfully employed L-tert-leucine as the transient chiral auxiliary to achieve the C−H alkynylation transformation with a broad range of heterocycles, e.g., pyrroles, thiophenes, and furans (Scheme 18)[71]. With this strategy, the Xie[72] and Liu[73] groups also independently realized the enantioselective formation of C−N and N−N axial frameworks.
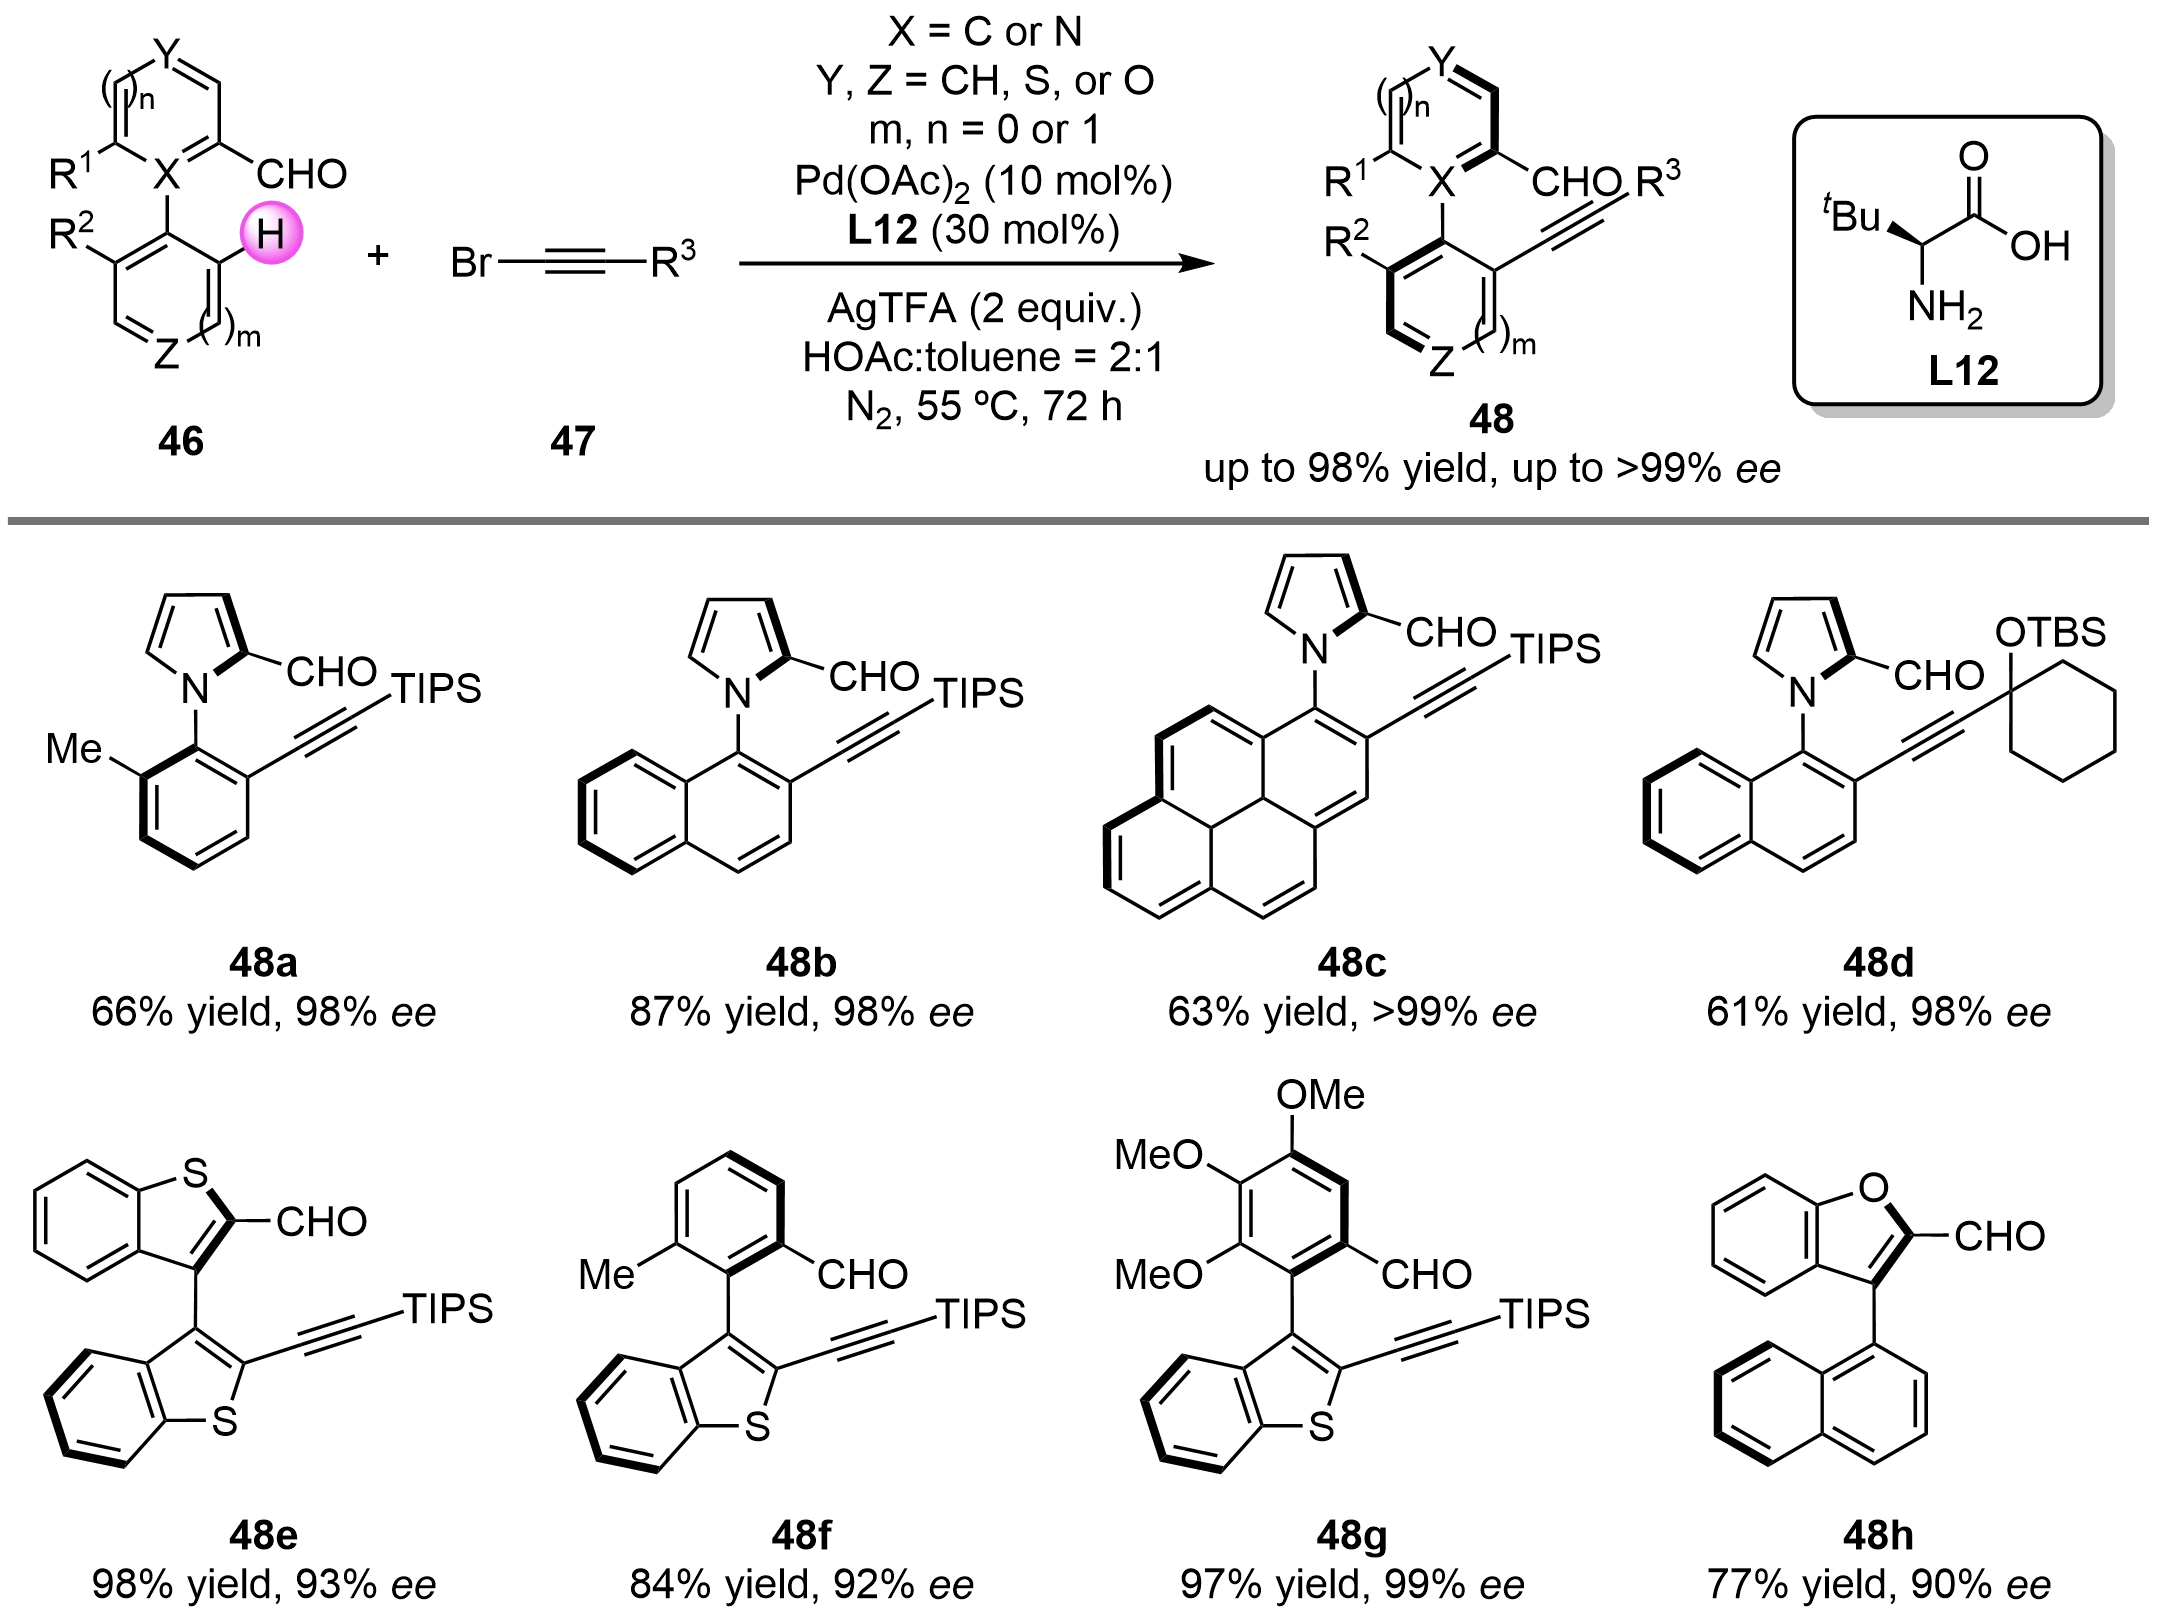
Scheme 18. Construction of C−N and C−C axial chirality by palladium-catalyzed asymmetric C−H alkynylation with a transient chiral auxiliary.
The C−H functionalization reactions with a transient directing group described above are all driven by chemical energy. The utilization of electrochemistry driven by sustainable and clean energy sources has also attracted considerable attention, especially targeting the synthesis of axially chiral biaryl compounds. In 2020, Ackermann and co-workers reported a palladium-catalyzed asymmetric C−H alkylation reaction by using a transient auxiliary, utilizing an electrochemical methodology to circumvent the need for stoichiometric amounts of oxidants. Under the optimized conditions, a series of axially chiral biaryls were obtained with excellent enantioselectivity in moderate to good yields (Scheme 19)[74].
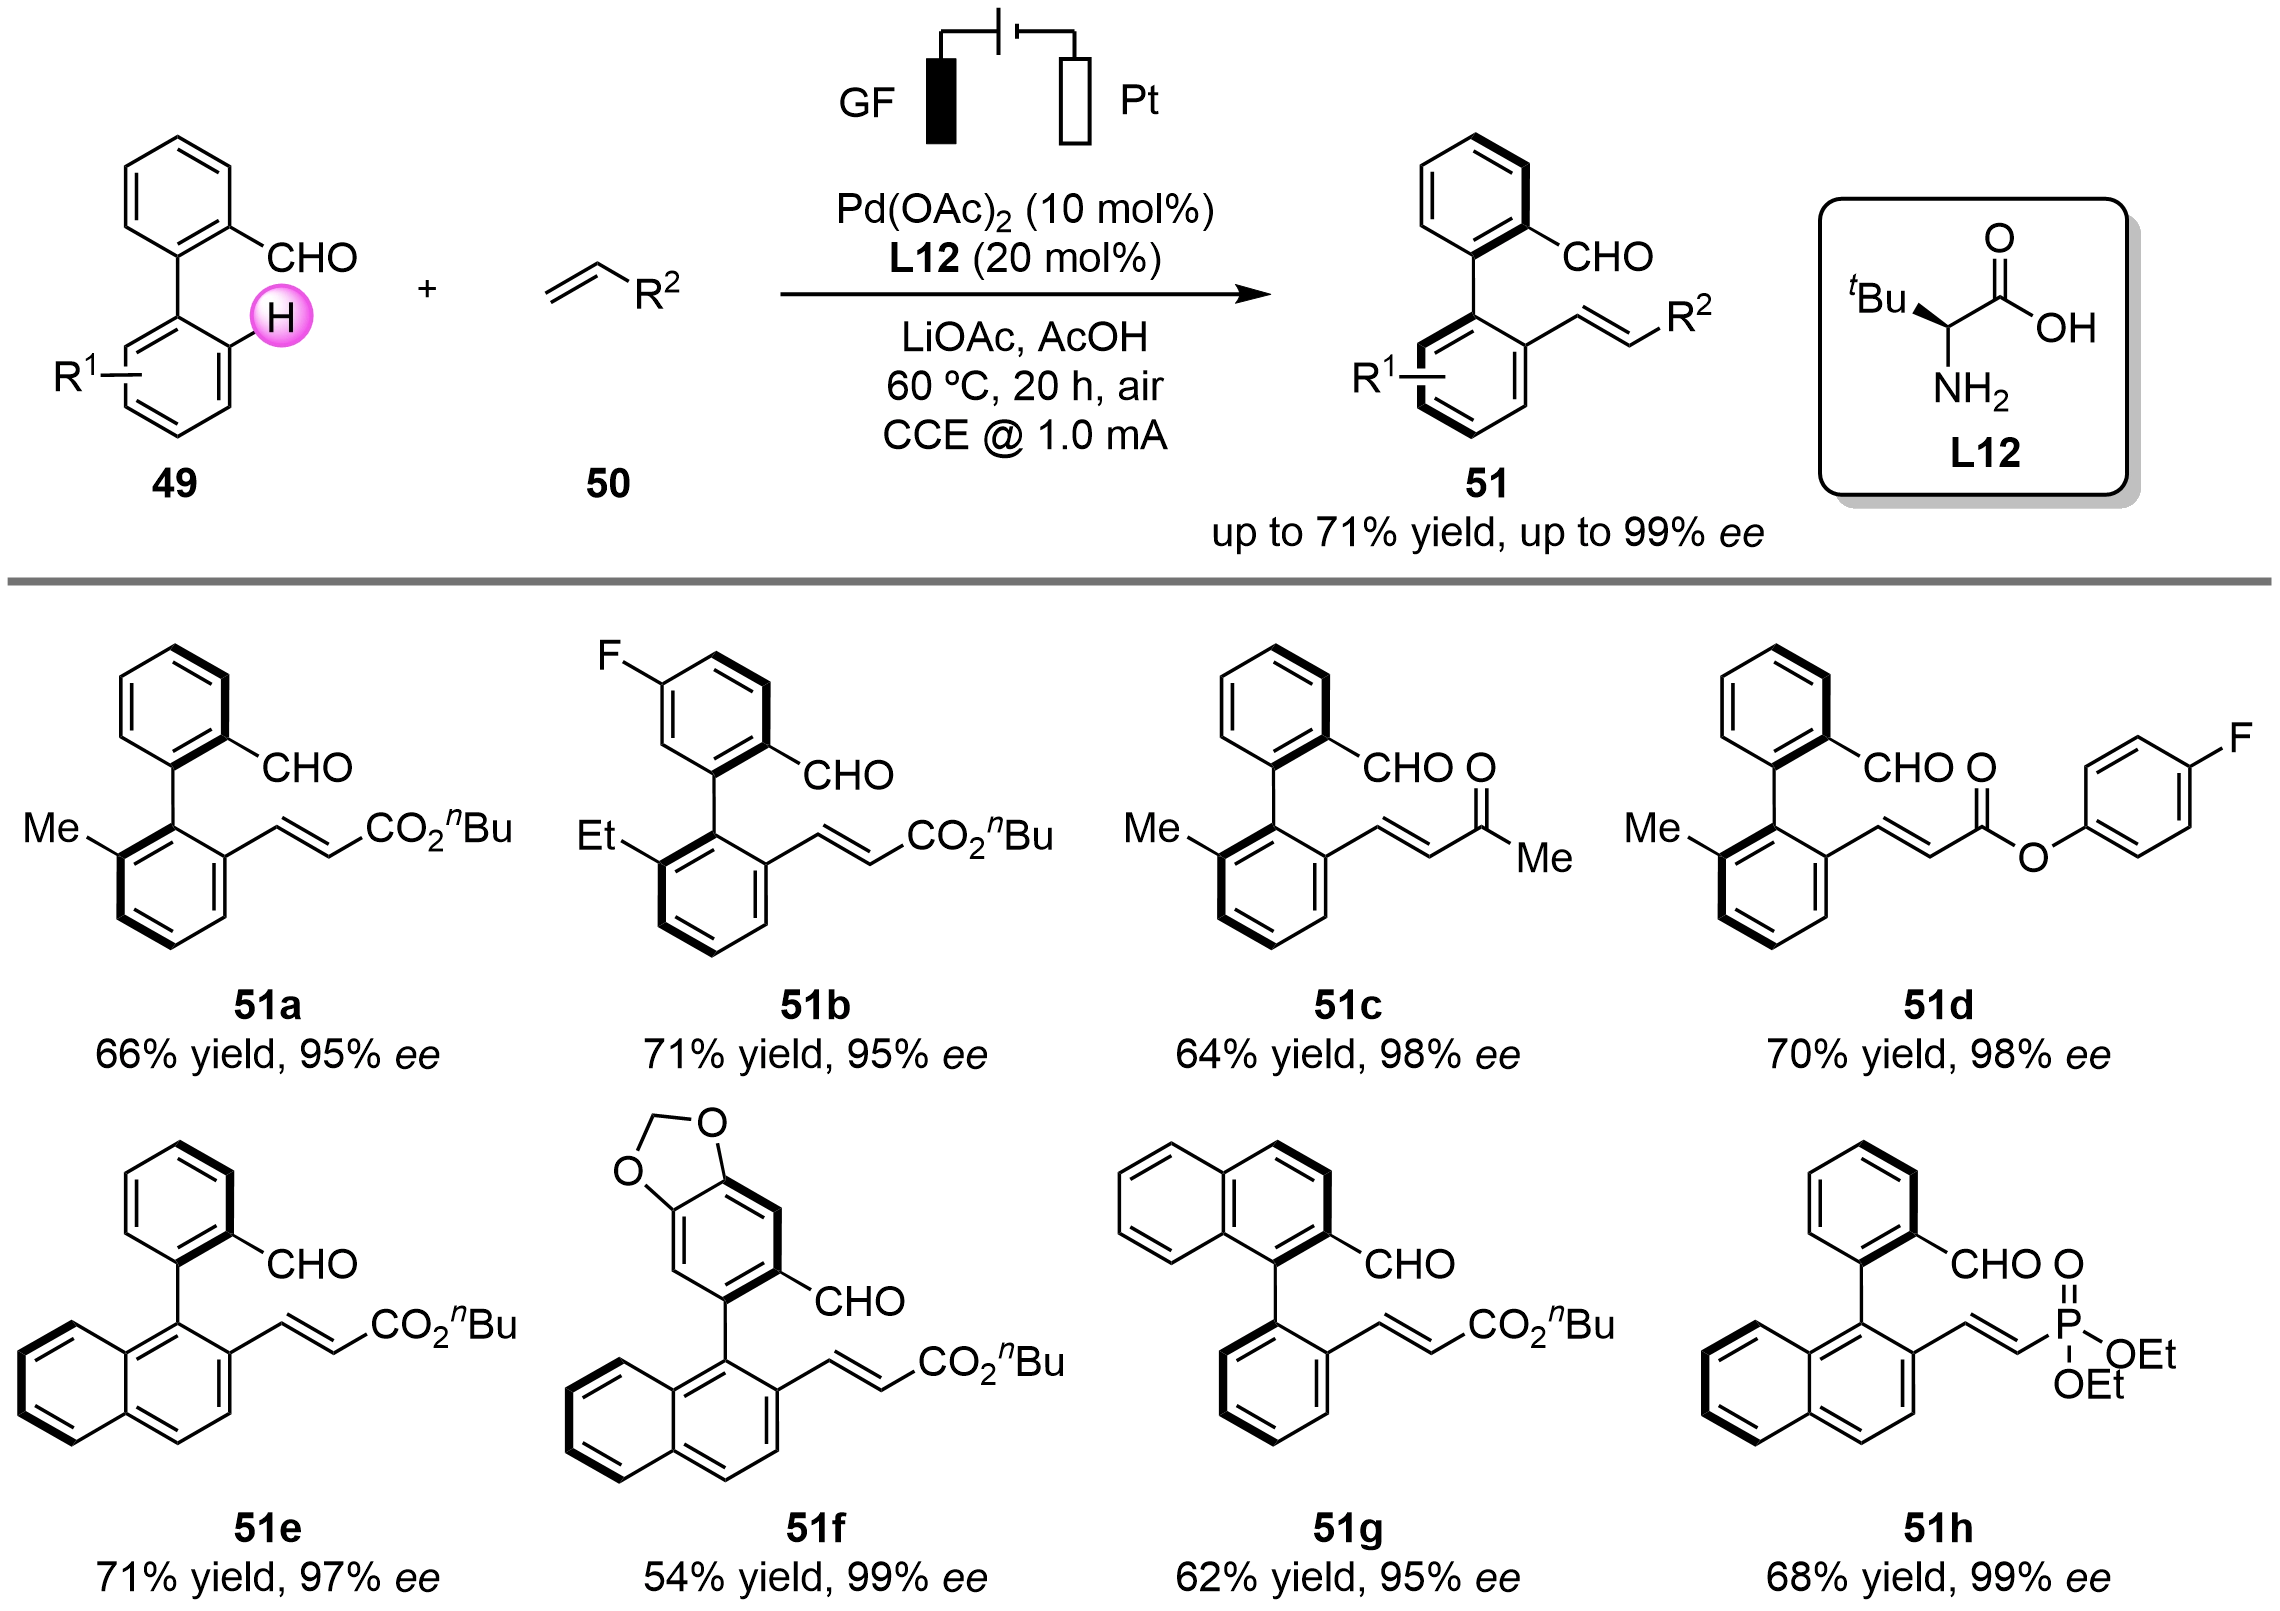
Scheme 19. Construction of C−C axial chirality by palladium-catalyzed electrochemical asymmetric C−H olefination.
2.3 DKR of biaryls with free axial rotation via directing group-free C−H functionalization
DKR of configurationally labile biaryls via DG-free C−H functionalization can be achieved by obviating the need for strongly coordinating directing groups or the pre-installation of a formyl group on the substrate scaffolds for a transient directing strategy. Instead, the incorporation of heteroatoms, such as nitrogen and oxygen, into the aromatic rings modulates the electronic properties, resulting in an uneven distribution of electron density that becomes significantly elevated in the proximity of the heteroatoms. Consequently, the presence of these heteroatoms enables precise control over the reaction enantioselectivity by leveraging steric hindrance alongside weak non-covalent interactions between the substrate and the chiral catalyst, including hydrogen bonding, π-π stacking, and C−H-π interactions (Scheme 20).
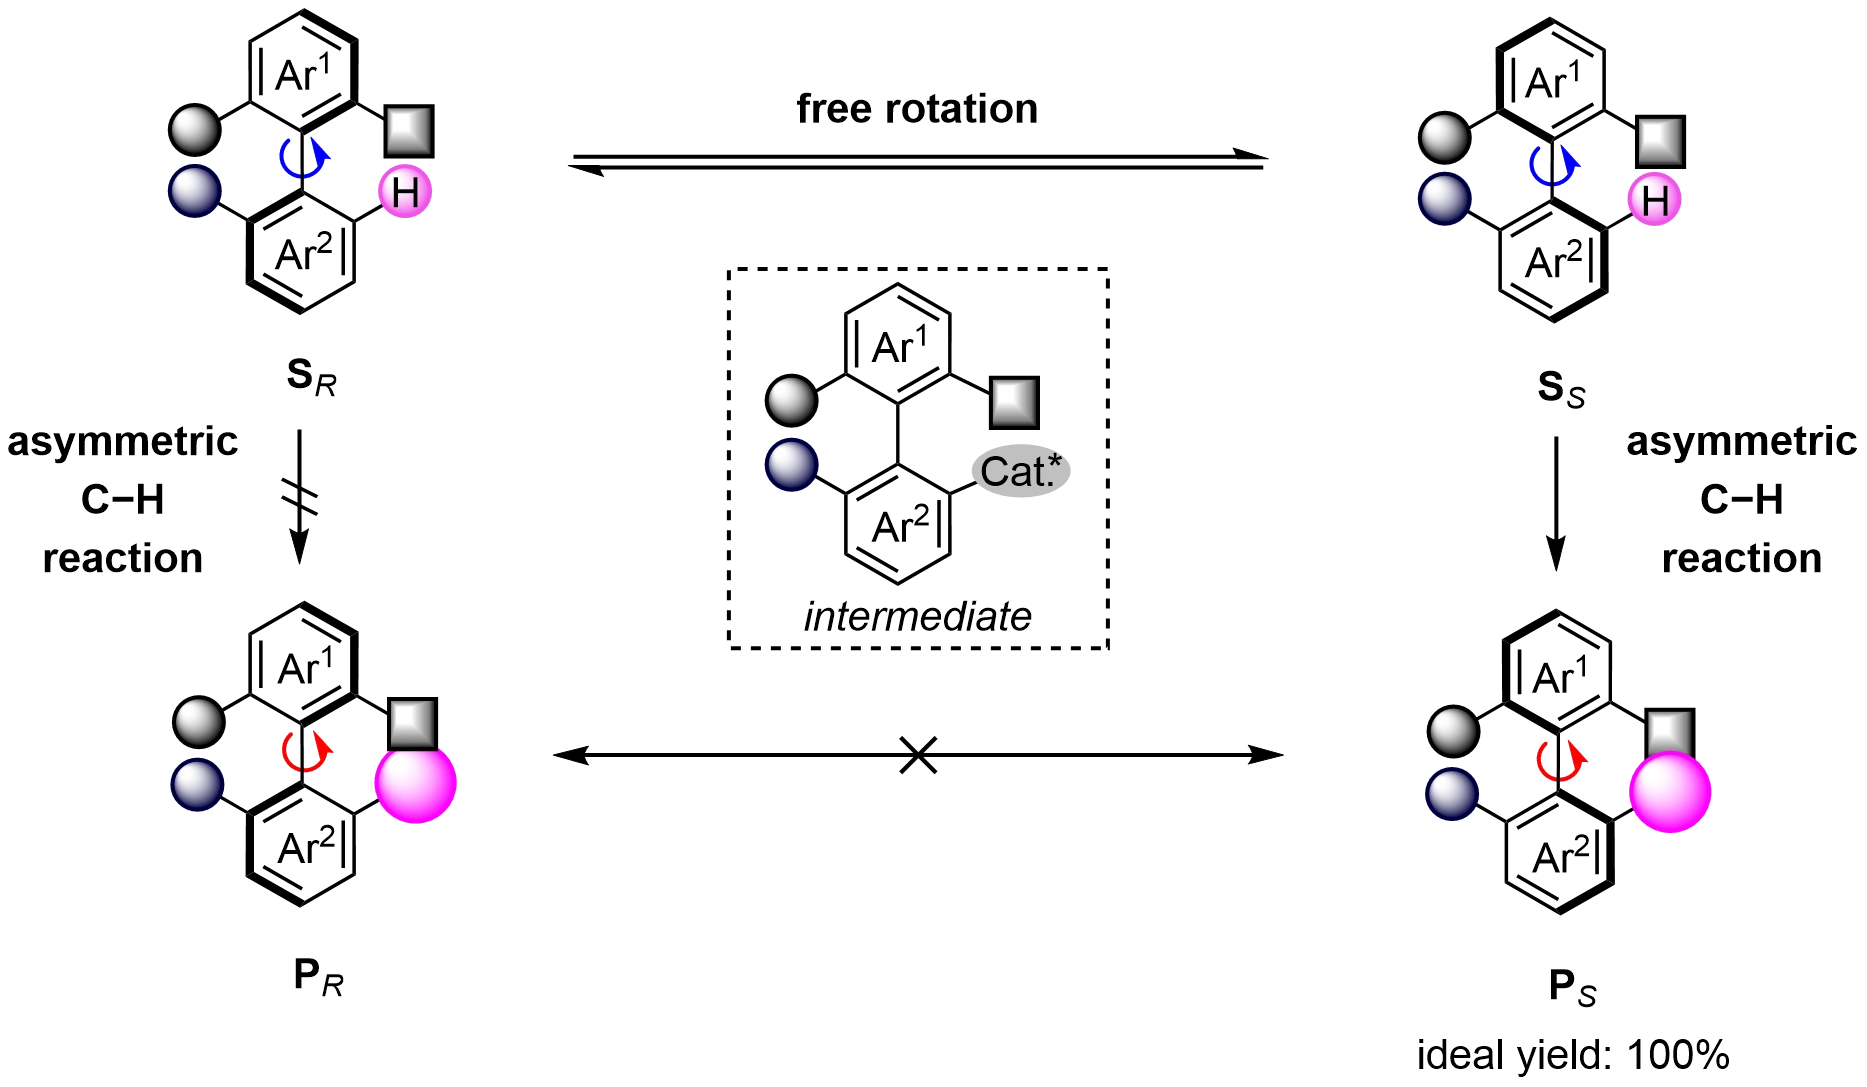
Scheme 20. DKR of biaryls with free axial rotation via DG-free C−H functionalization. DKR: dynamic kinetic resolution; DG: directing group.
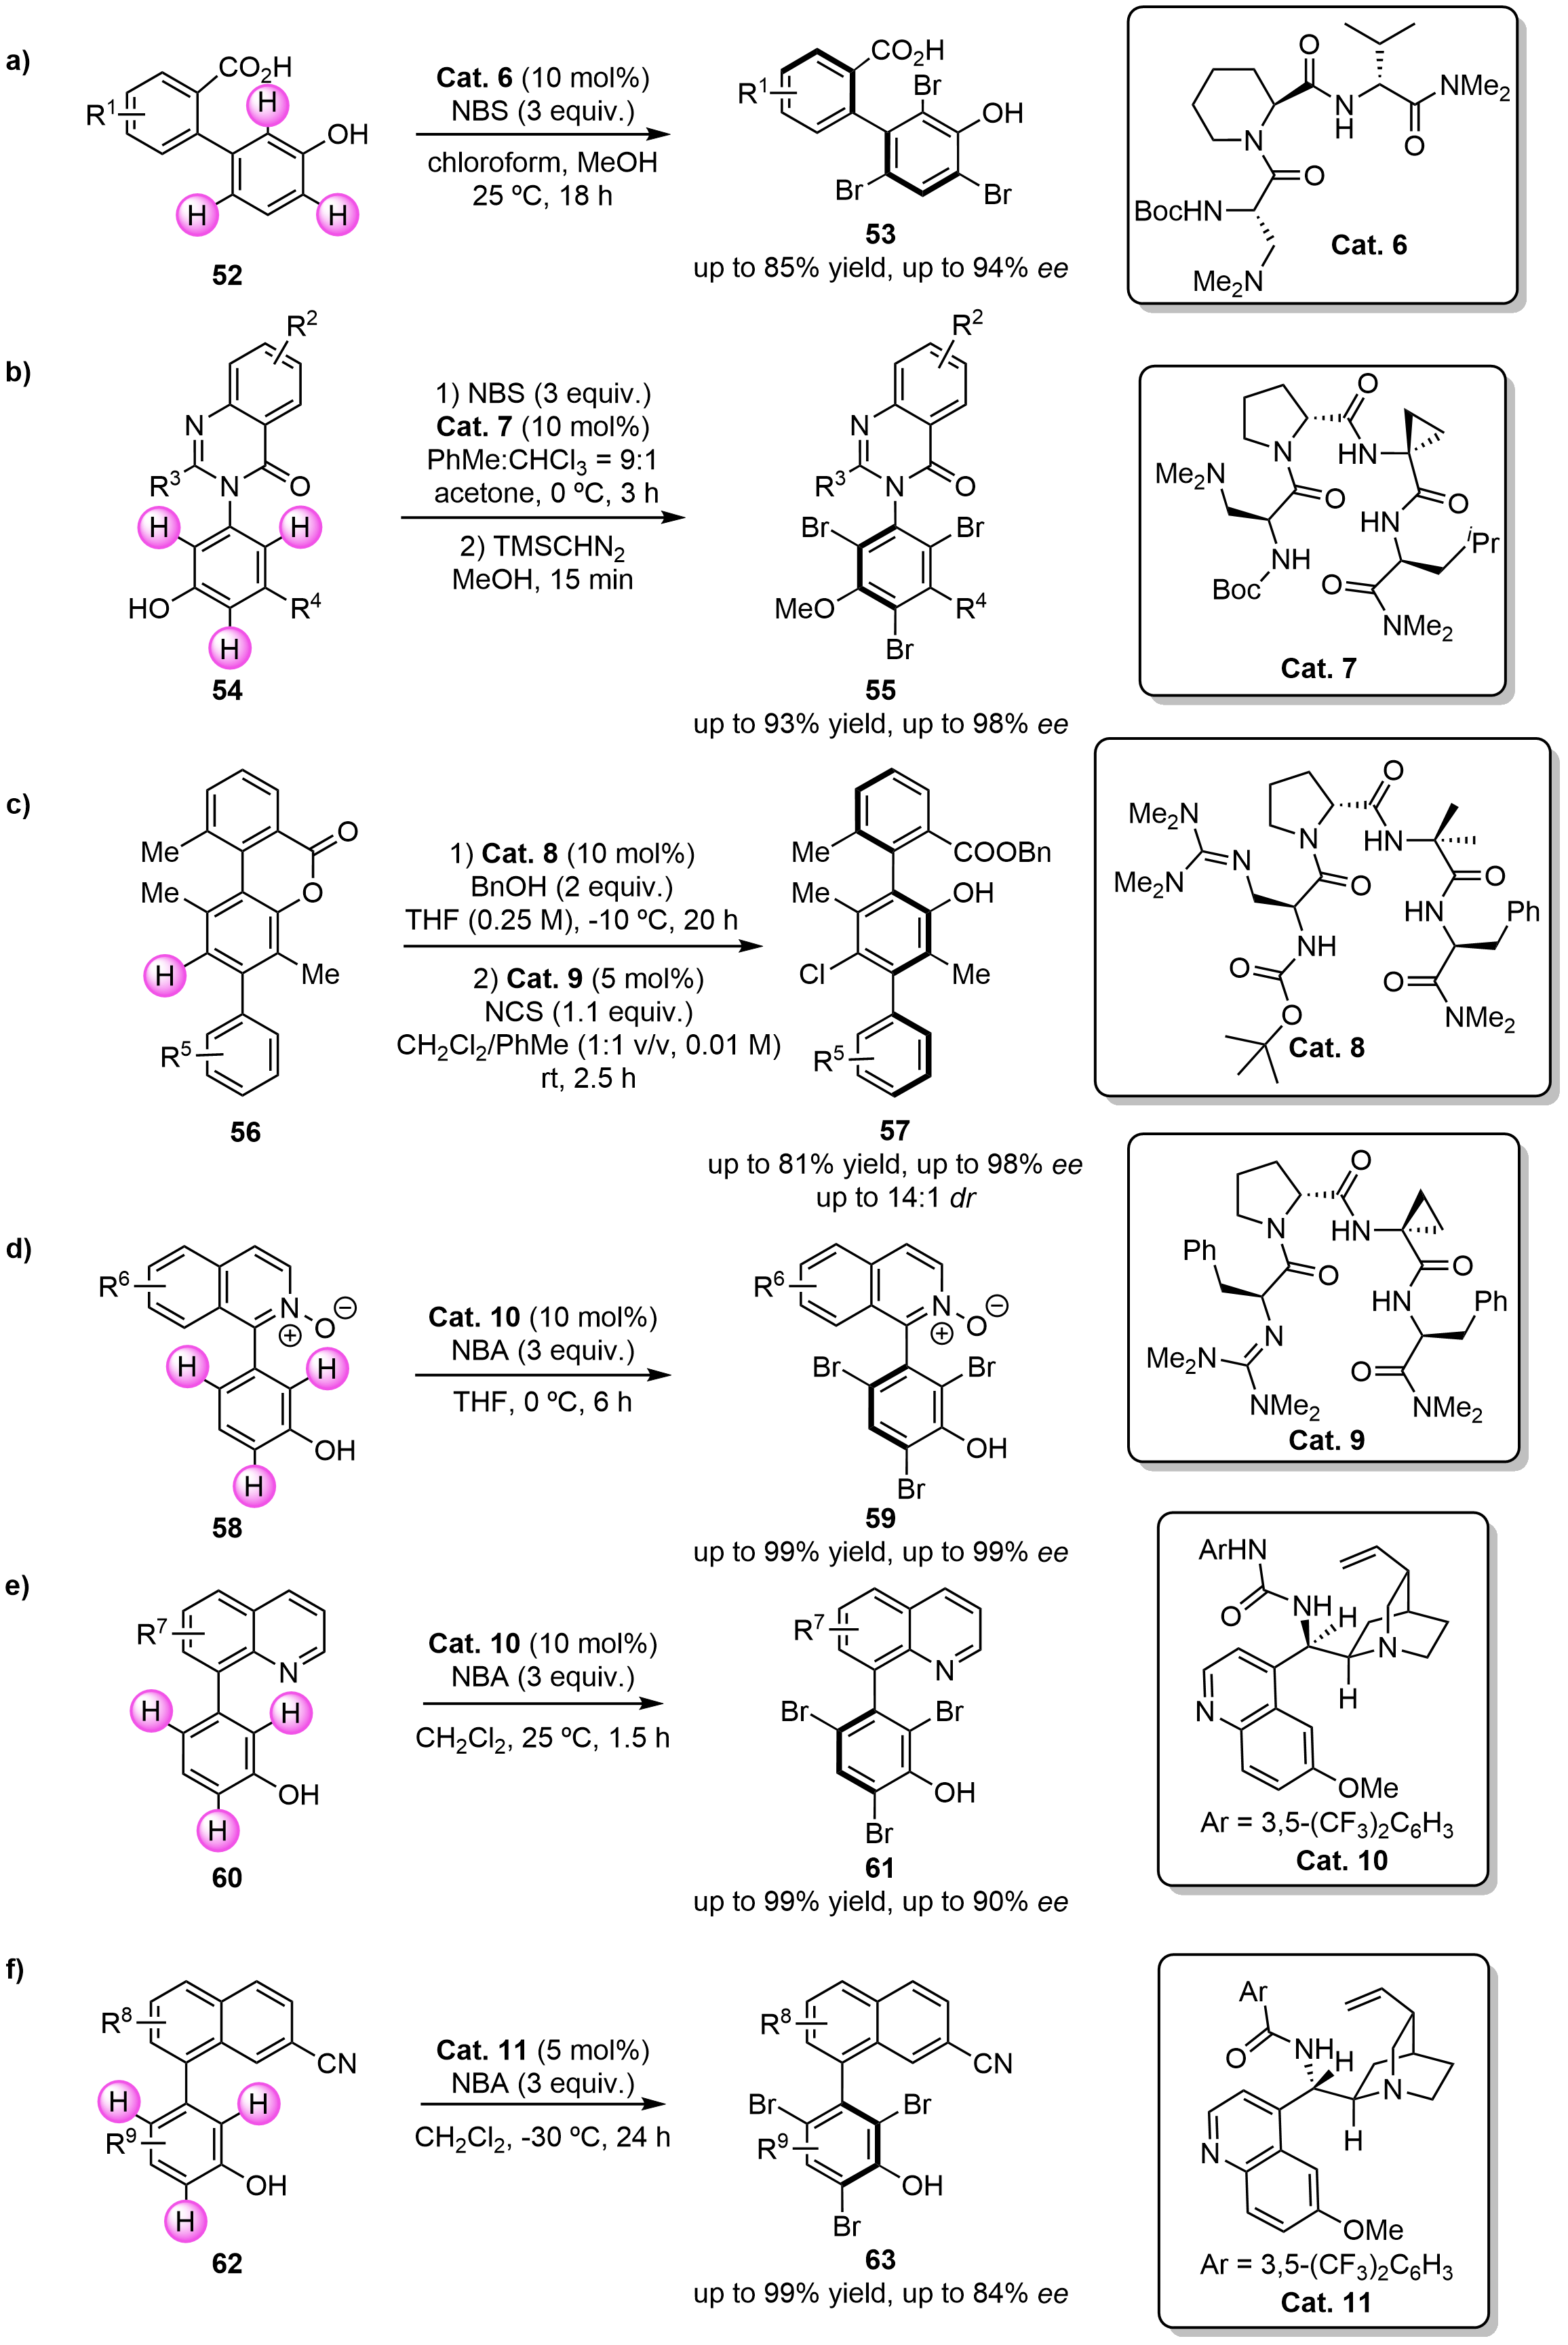
In 2010, Miller and co-workers reported a DKR of biaryl atropisomers via peptide-catalyzed asymmetric bromination (Scheme 21a)[75]. Notably, 3-arylphenol substrate 52 was observed to undergo free rotation, as well as its partially brominated product (e.g., mono- and o,p-dibrominated) until the two ortho-bromo substituents were both installed. In this way, a higher energy barrier was able to prevent the axial rotation in tribrominated products. The hydrogen bond between the phenolic proton and the amide group in the chiral organocatalyst blocks axial bond rotation at the stage of the hydrogen-bond-linked intermediate. Therefore, under the catalysis of peptide Cat. 6, tribrominated product 53 was produced in high yield (up to 85%) with high enantioselectivity (up to 94% ee). In 2015, the same research group subsequently established peptide-catalyzed asymmetric C−H bromination of N-aryl quinazolinones for the construction of axially chiral quinazolinones 55 (Scheme 21b)[76]. The results of DFT calculations showed that the substrates generally bear an aryl-aryl axis of free rotation. However, upon introducing bromine atoms at both ortho-positions of the C−N bond, a significant increase in the rotational energy barrier was observed, ensuring the stability of enantioenriched products. Further, the authors combined the enantioselective ring-opening of lactones with electrophilic chlorine/bromine substitution to accomplish the synthesis of two-axis terphenyl atropisomers in two steps (Scheme 21c)[77]. Similarly, the Matsubara group employed a quinine catalyst to achieve the DKR of biaryl compounds through electrophilic halogenation (Scheme 21d,e,f)[78-80].
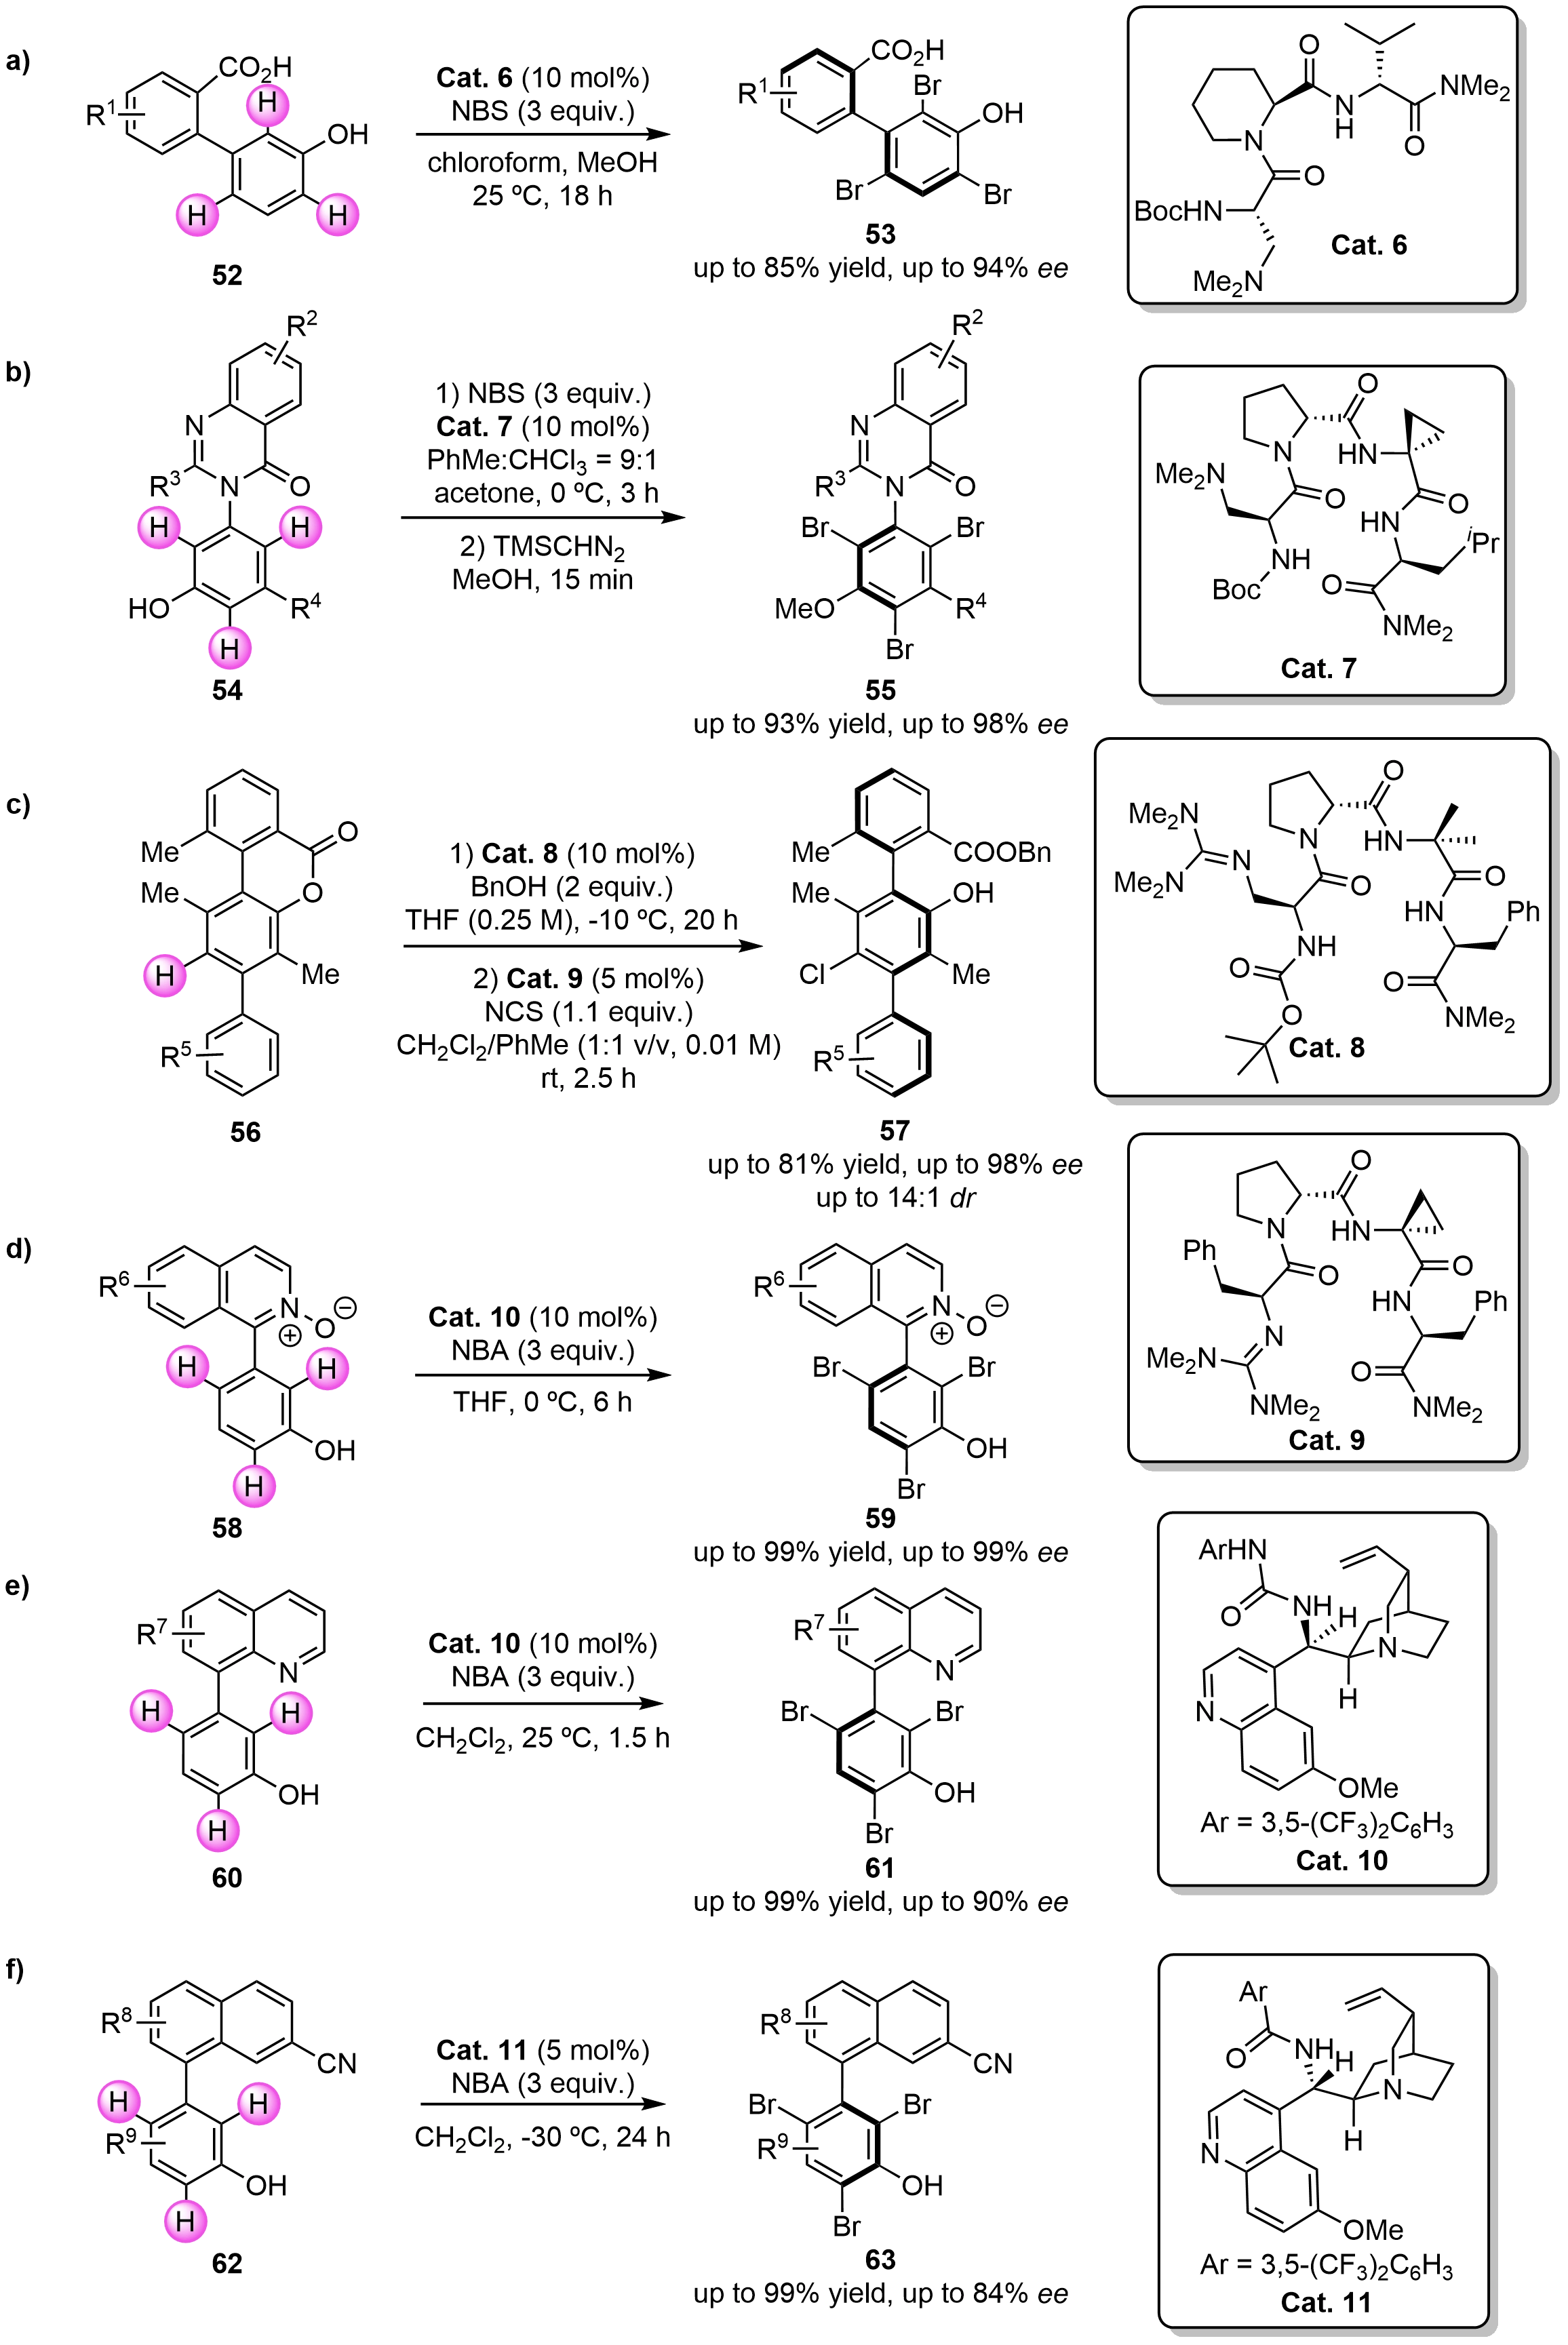
Scheme 21. DKR of biaryl atropisomers via peptide-catalyzed asymmetric halogenation.
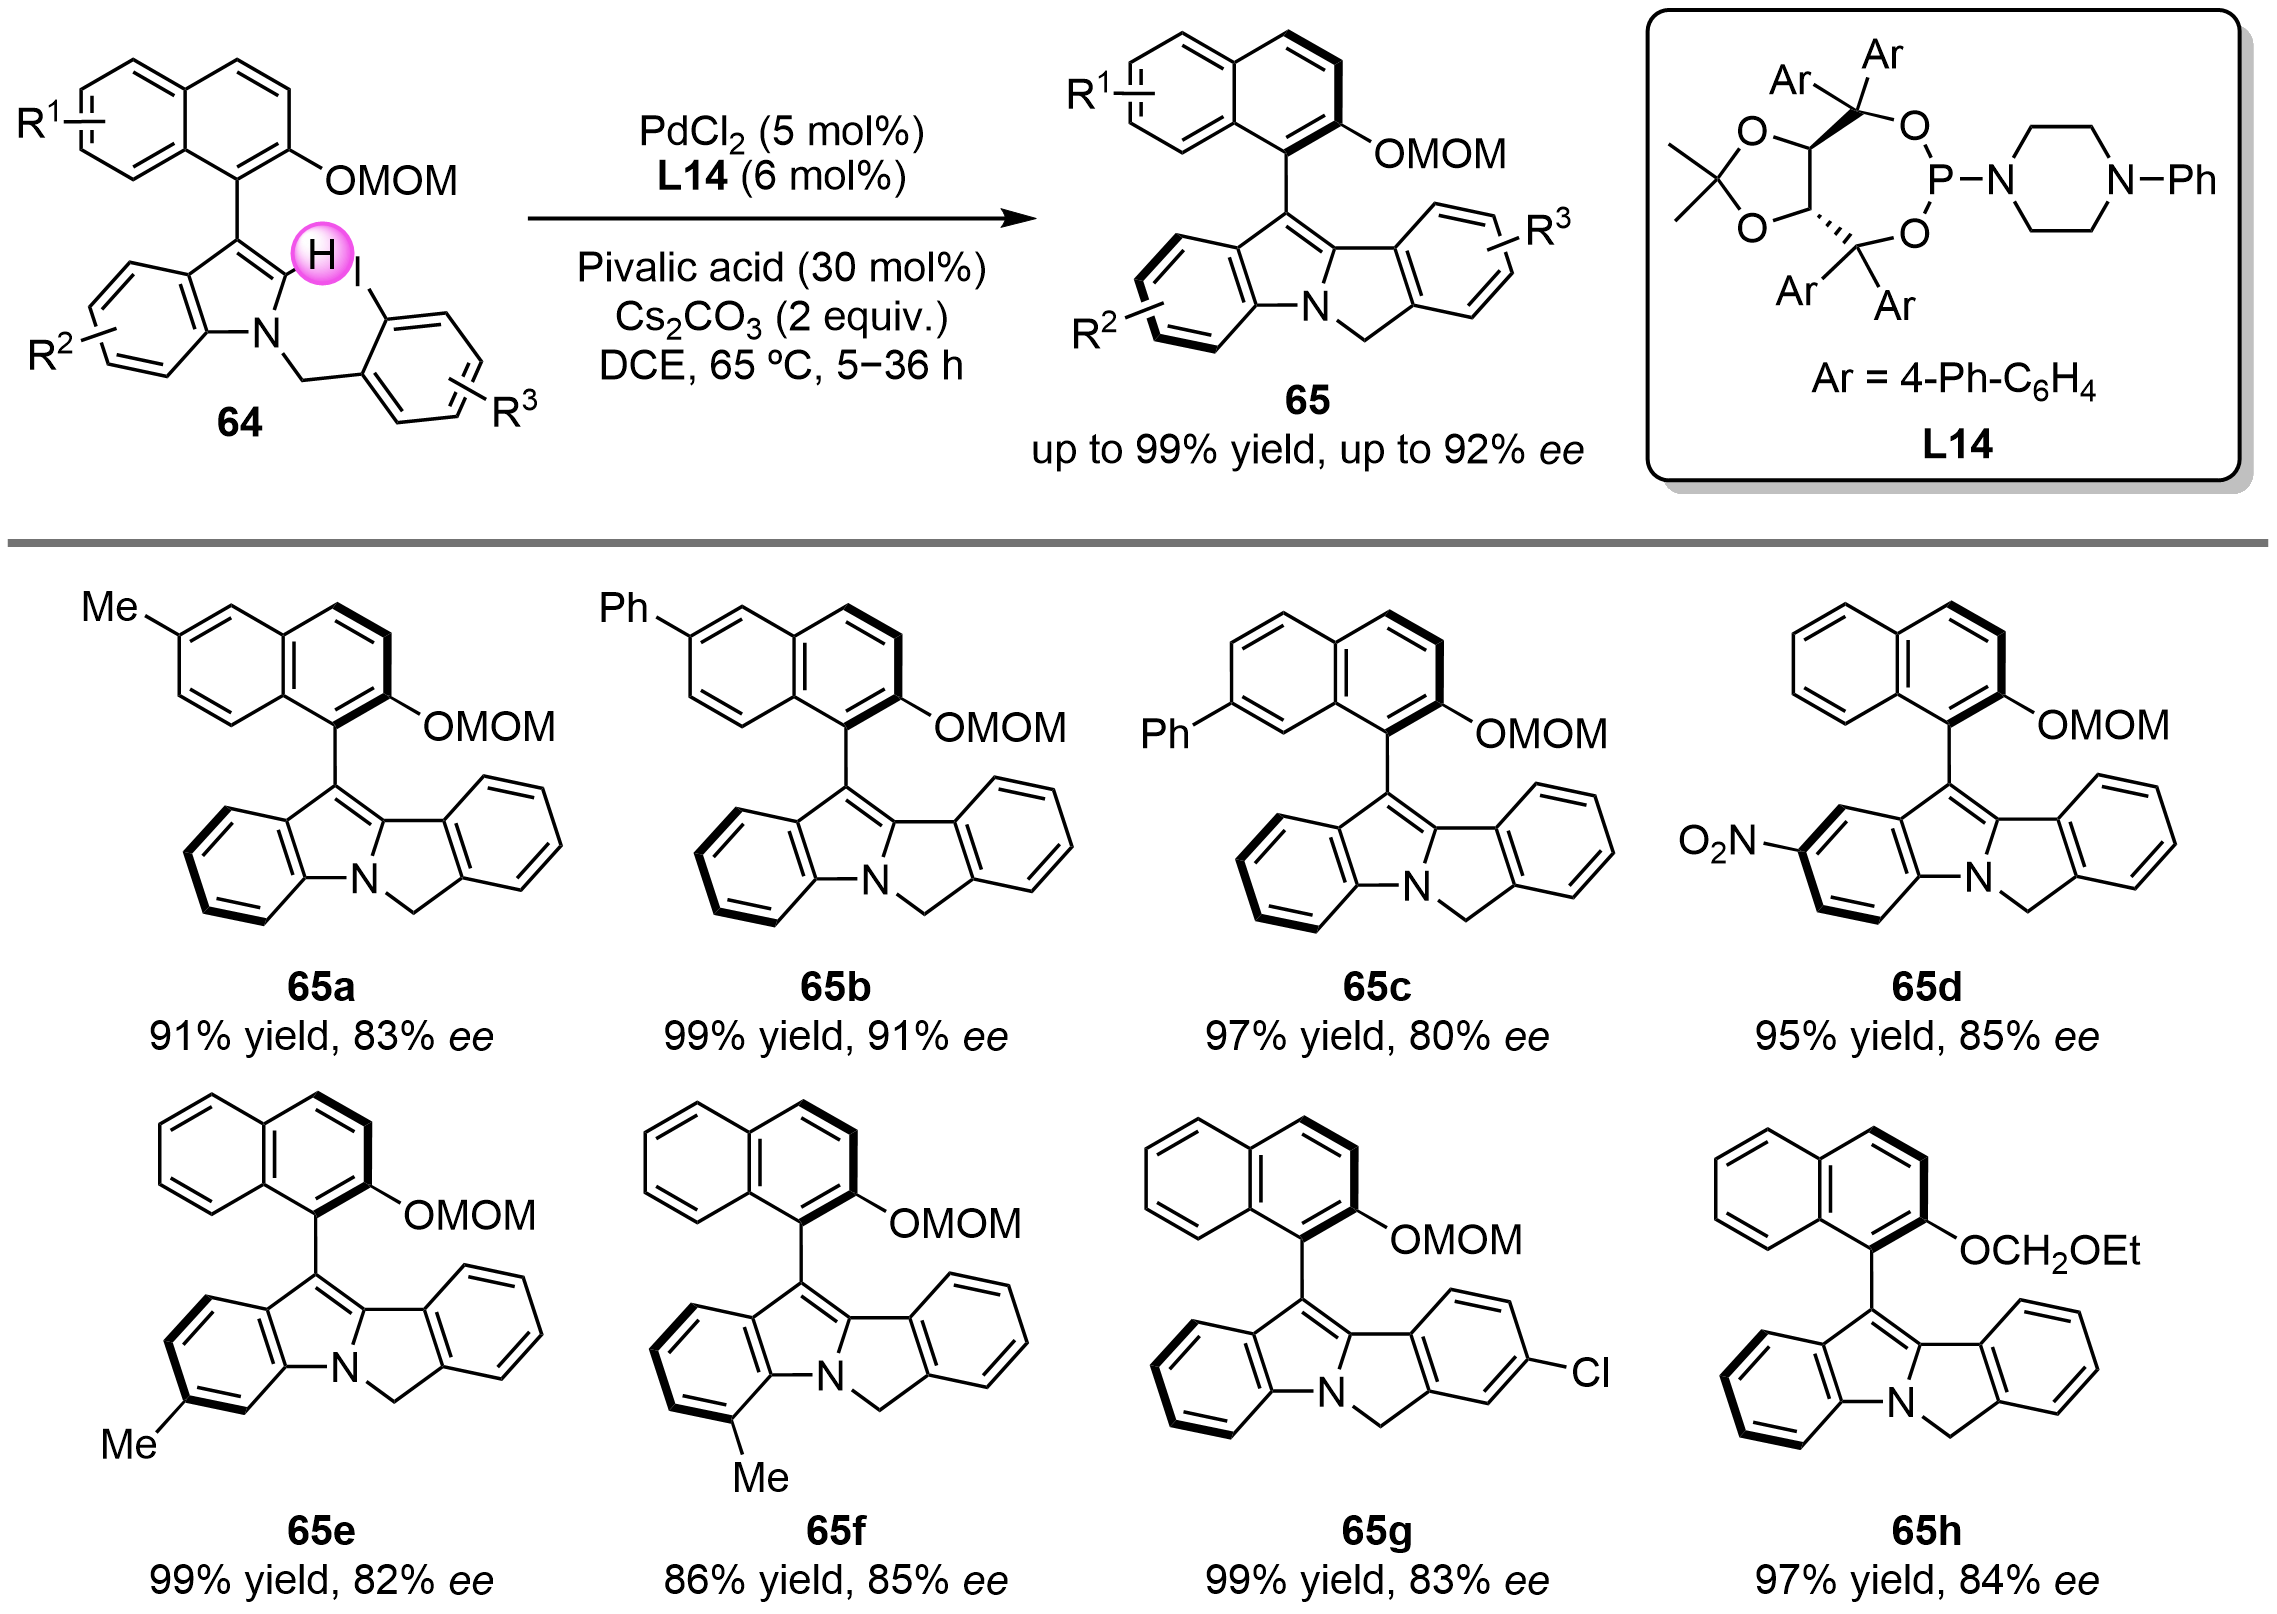
In 2017, the Gu group employed a catalytic system comprising PdCl2 and TADDOL-derived phosphoramide ligand (L14) to accomplish the asymmetric intramolecular C−H arylation of 3-arylindole derivatives. Notably, the overall C−H arylation transformations were realized without the involvement of a directing group. Axially chiral 3-arylindoles bearing either an electron-withdrawing group (EWG), or an electron-donating group (EDG), were easily accessible in excellent yield with decent enantioselectivity (Scheme 22)[81].
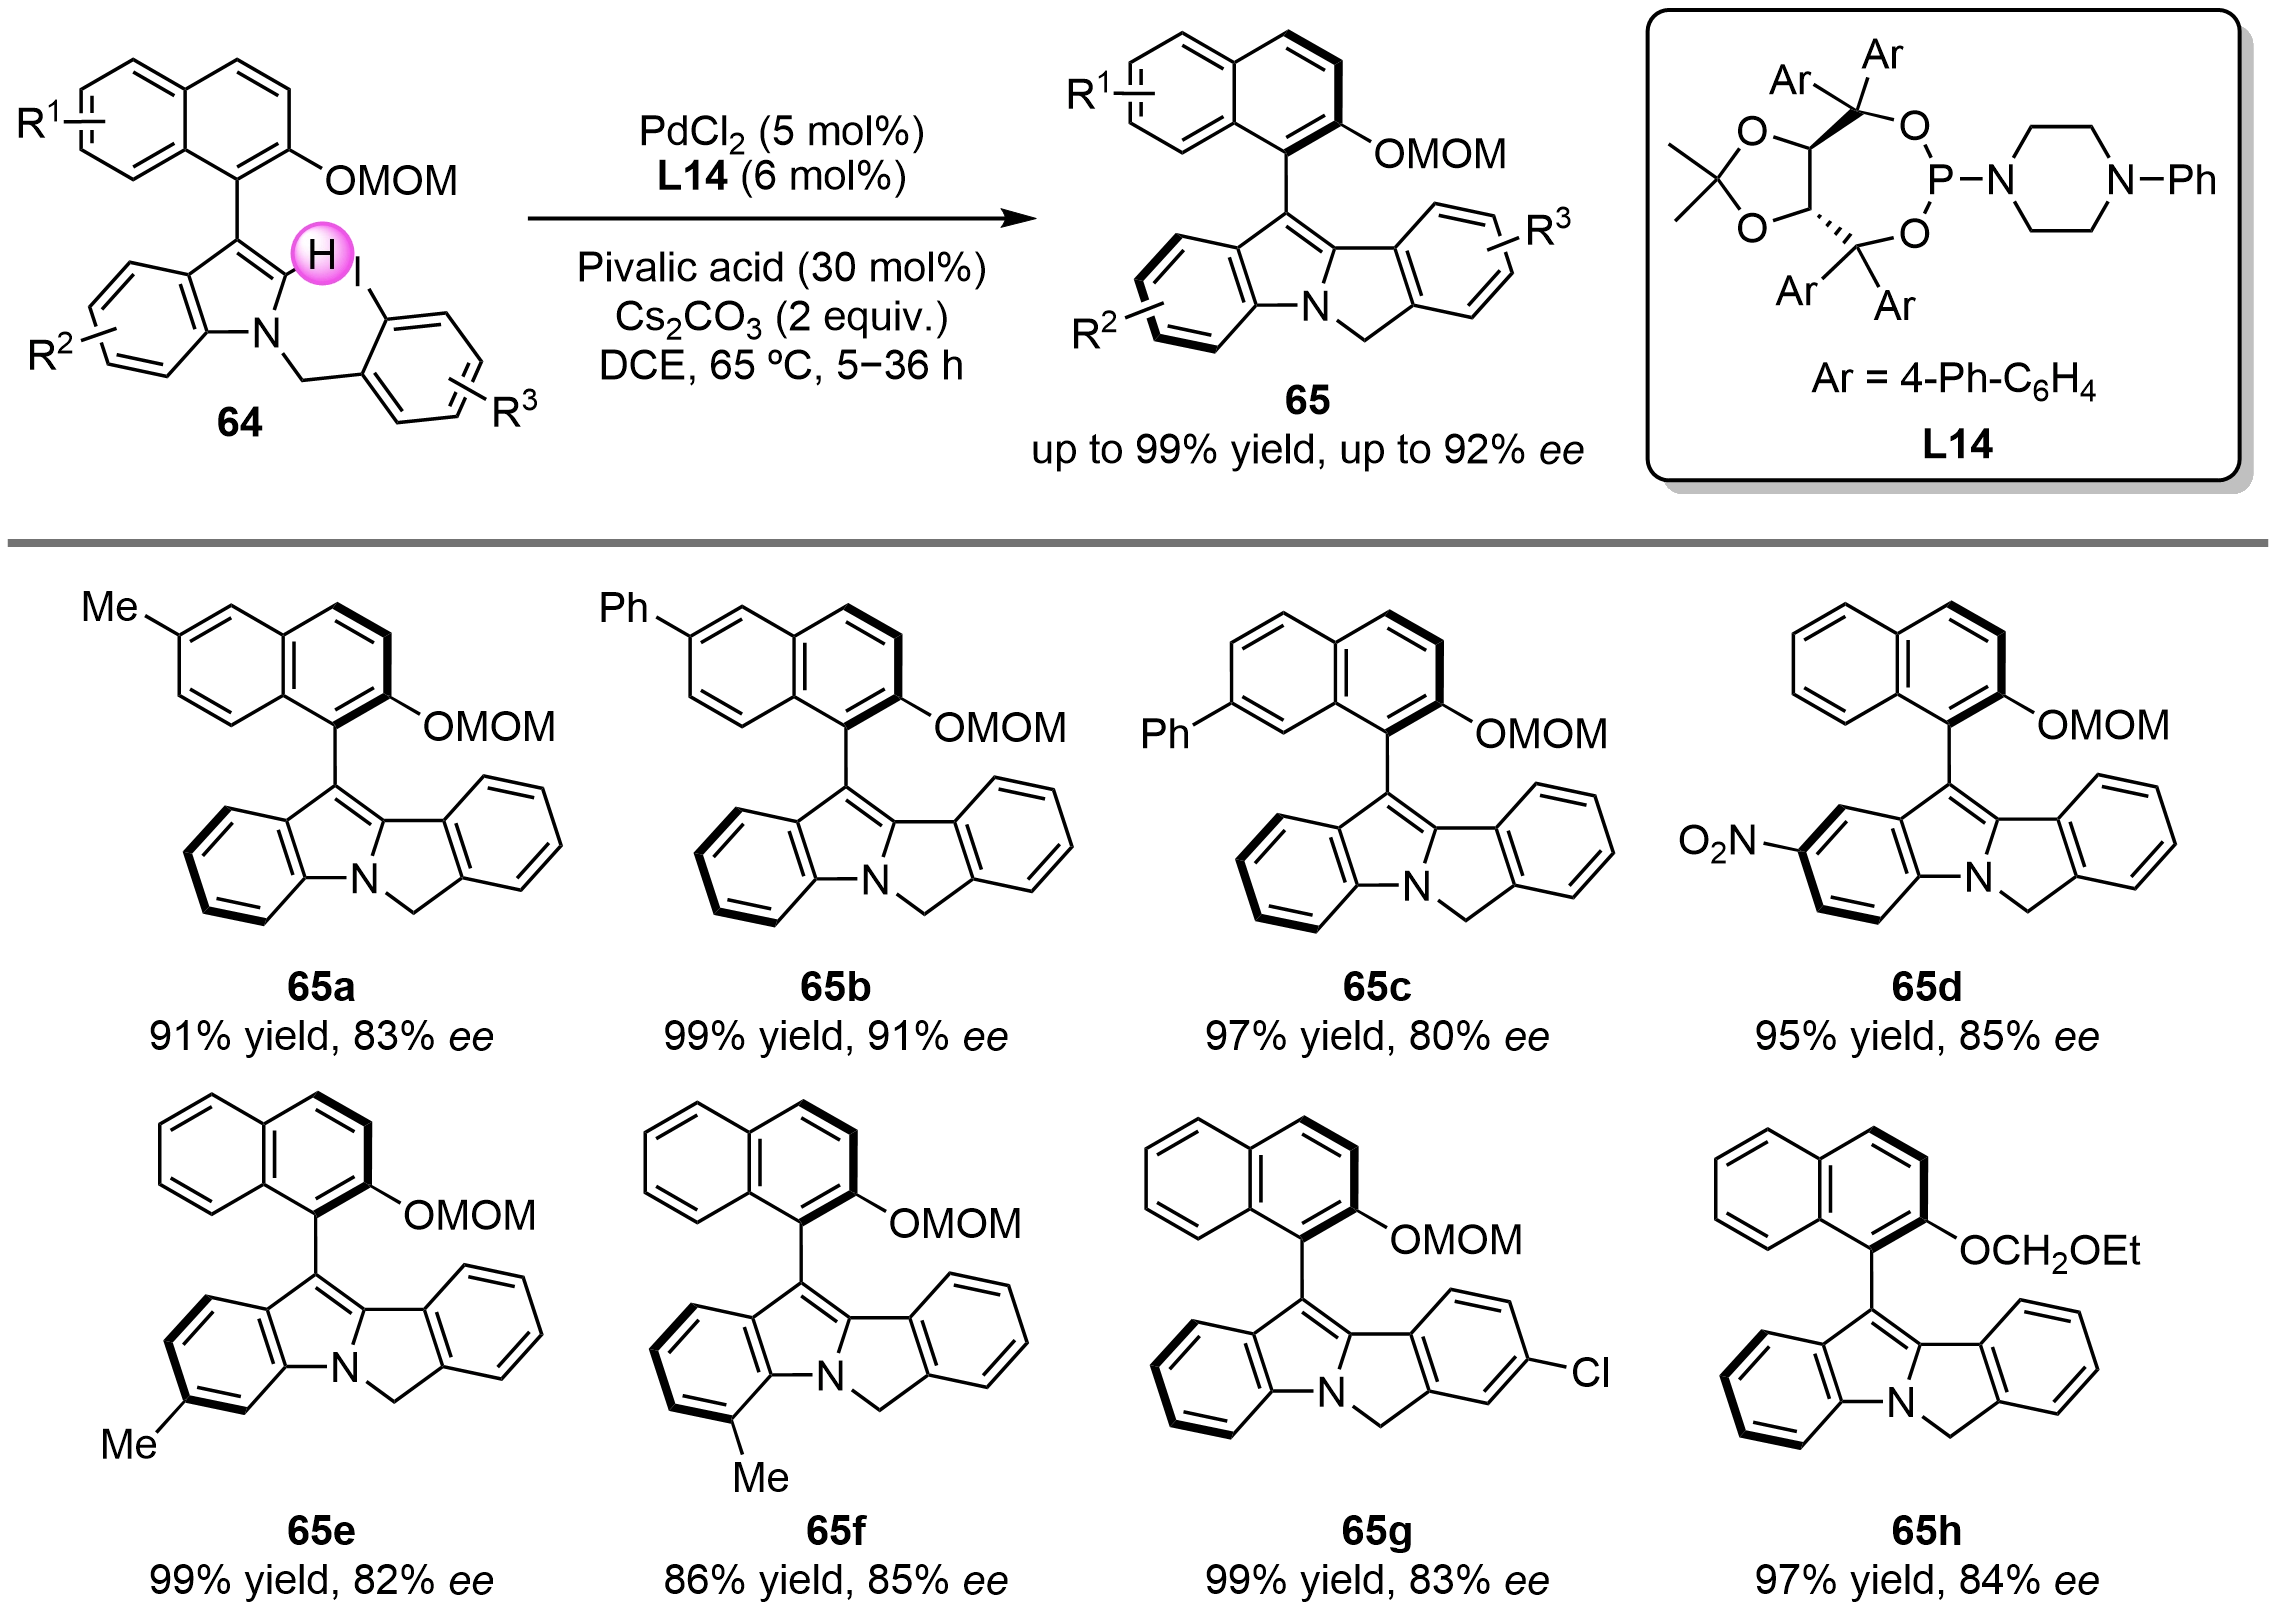
Scheme 22. Construction of C−C axial chirality by DG-free palladium-catalyzed asymmetric C−H annulation. DG: directing group.
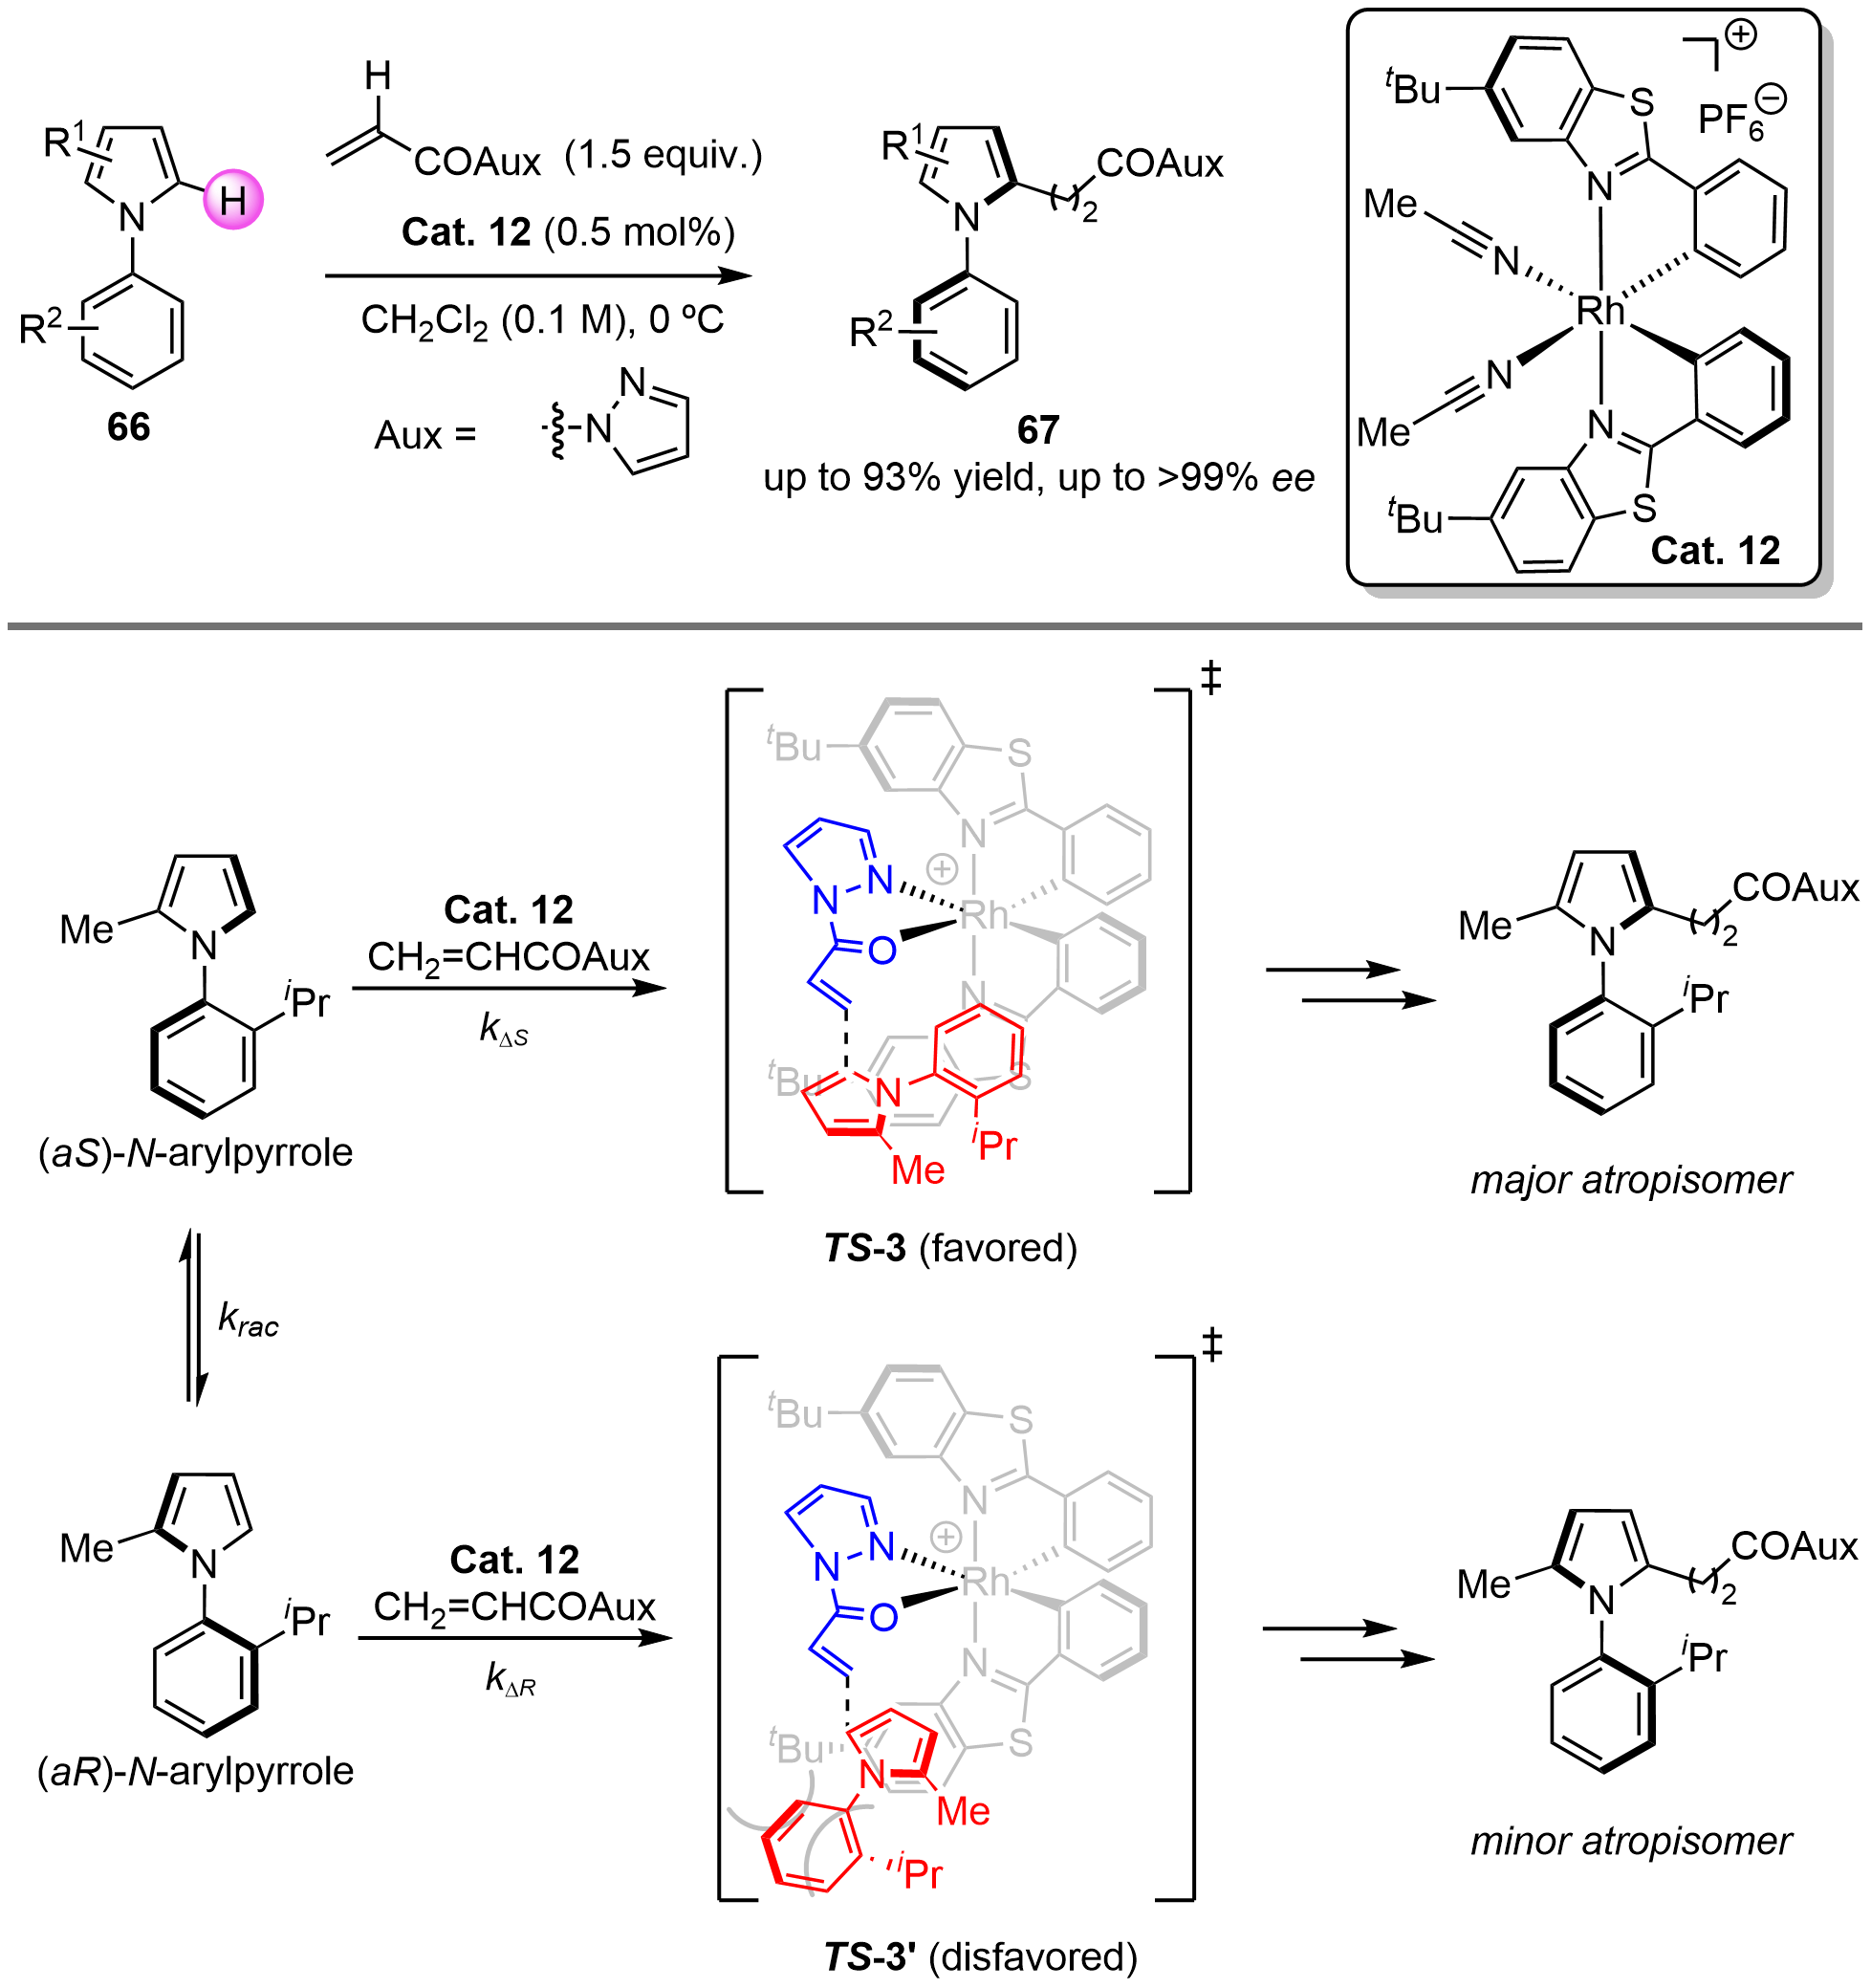
In 2020, the Meggers group exploited a unique chiral-at-metal rhodium catalyst (Cat. 12) to promote the asymmetric C−H alkylation of N-arylpyrroles with alkenyl amides in excellent yield (up to 93% yield) with high enantioselectivity (up to >99% ee). Under standard conditions, N-arylpyrrole substrates 66 exist as transient atropisomers that undergo rapid racemization. However, by introducing an electrophile at the highly nucleophilic α-position of the pyrrole ring, the rotational barrier around the C−N axis can be effectively hindered. This covalent functionalization successfully locks the biaryl scaffold, thereby establishing a configurationally stable axial chirality during the transformation. To demonstrate the origins of the stereoselectivity, they also conducted DFT calculations on the reactions. The Cat. 12 activated acrylpyrazole can react with both atropisomers. The reaction barrier of the (aS)-N-arylpyrrole (TS-3) is 2.1 kcal/mol lower than that of the (aR)-N-arylpyrrole (TS-3'). The isopropyl group on the N-arylpyrroles undergoes steric repulsion with the tert-butyl group of the ligand (closest H···H distance is 2.35 Å) in TS-3', while no unstable interaction occurs in TS-3. This match-mismatch effect in the transition states led to the stereoselectivity (Scheme 23)[82].
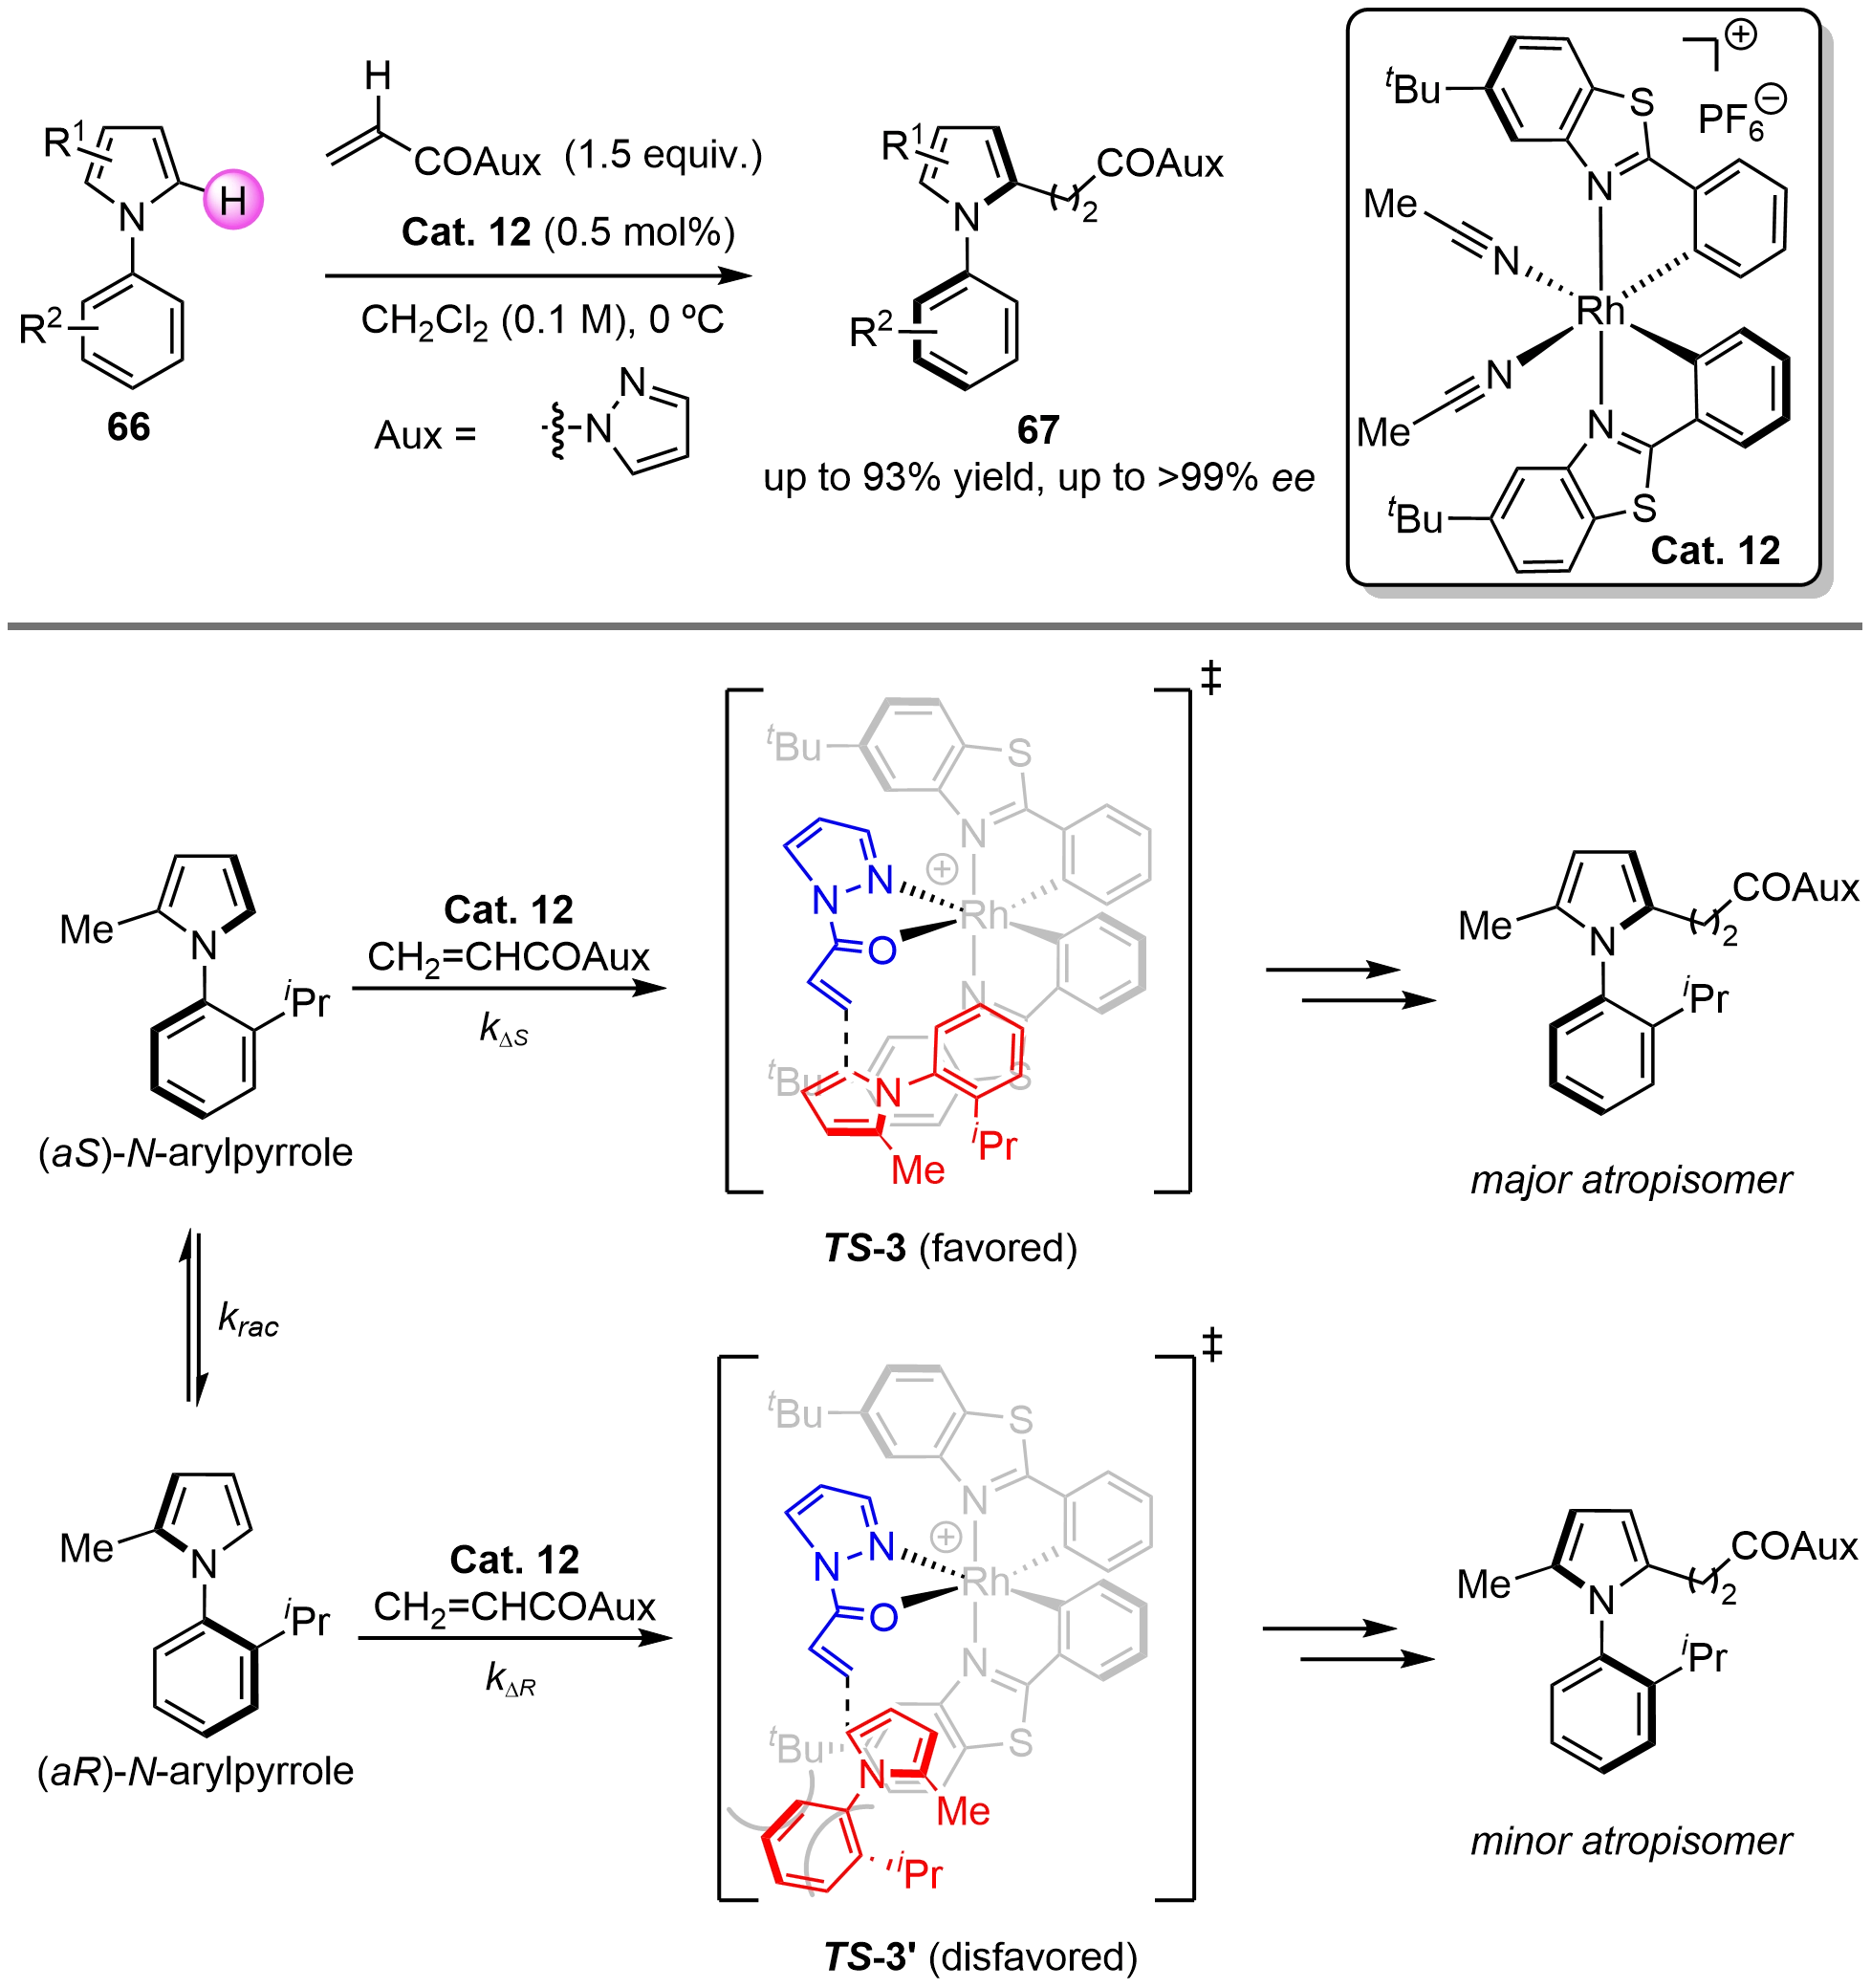
Scheme 23. Construction of C−N axial chirality catalyzed by a chiral-at-metal rhodium complex.
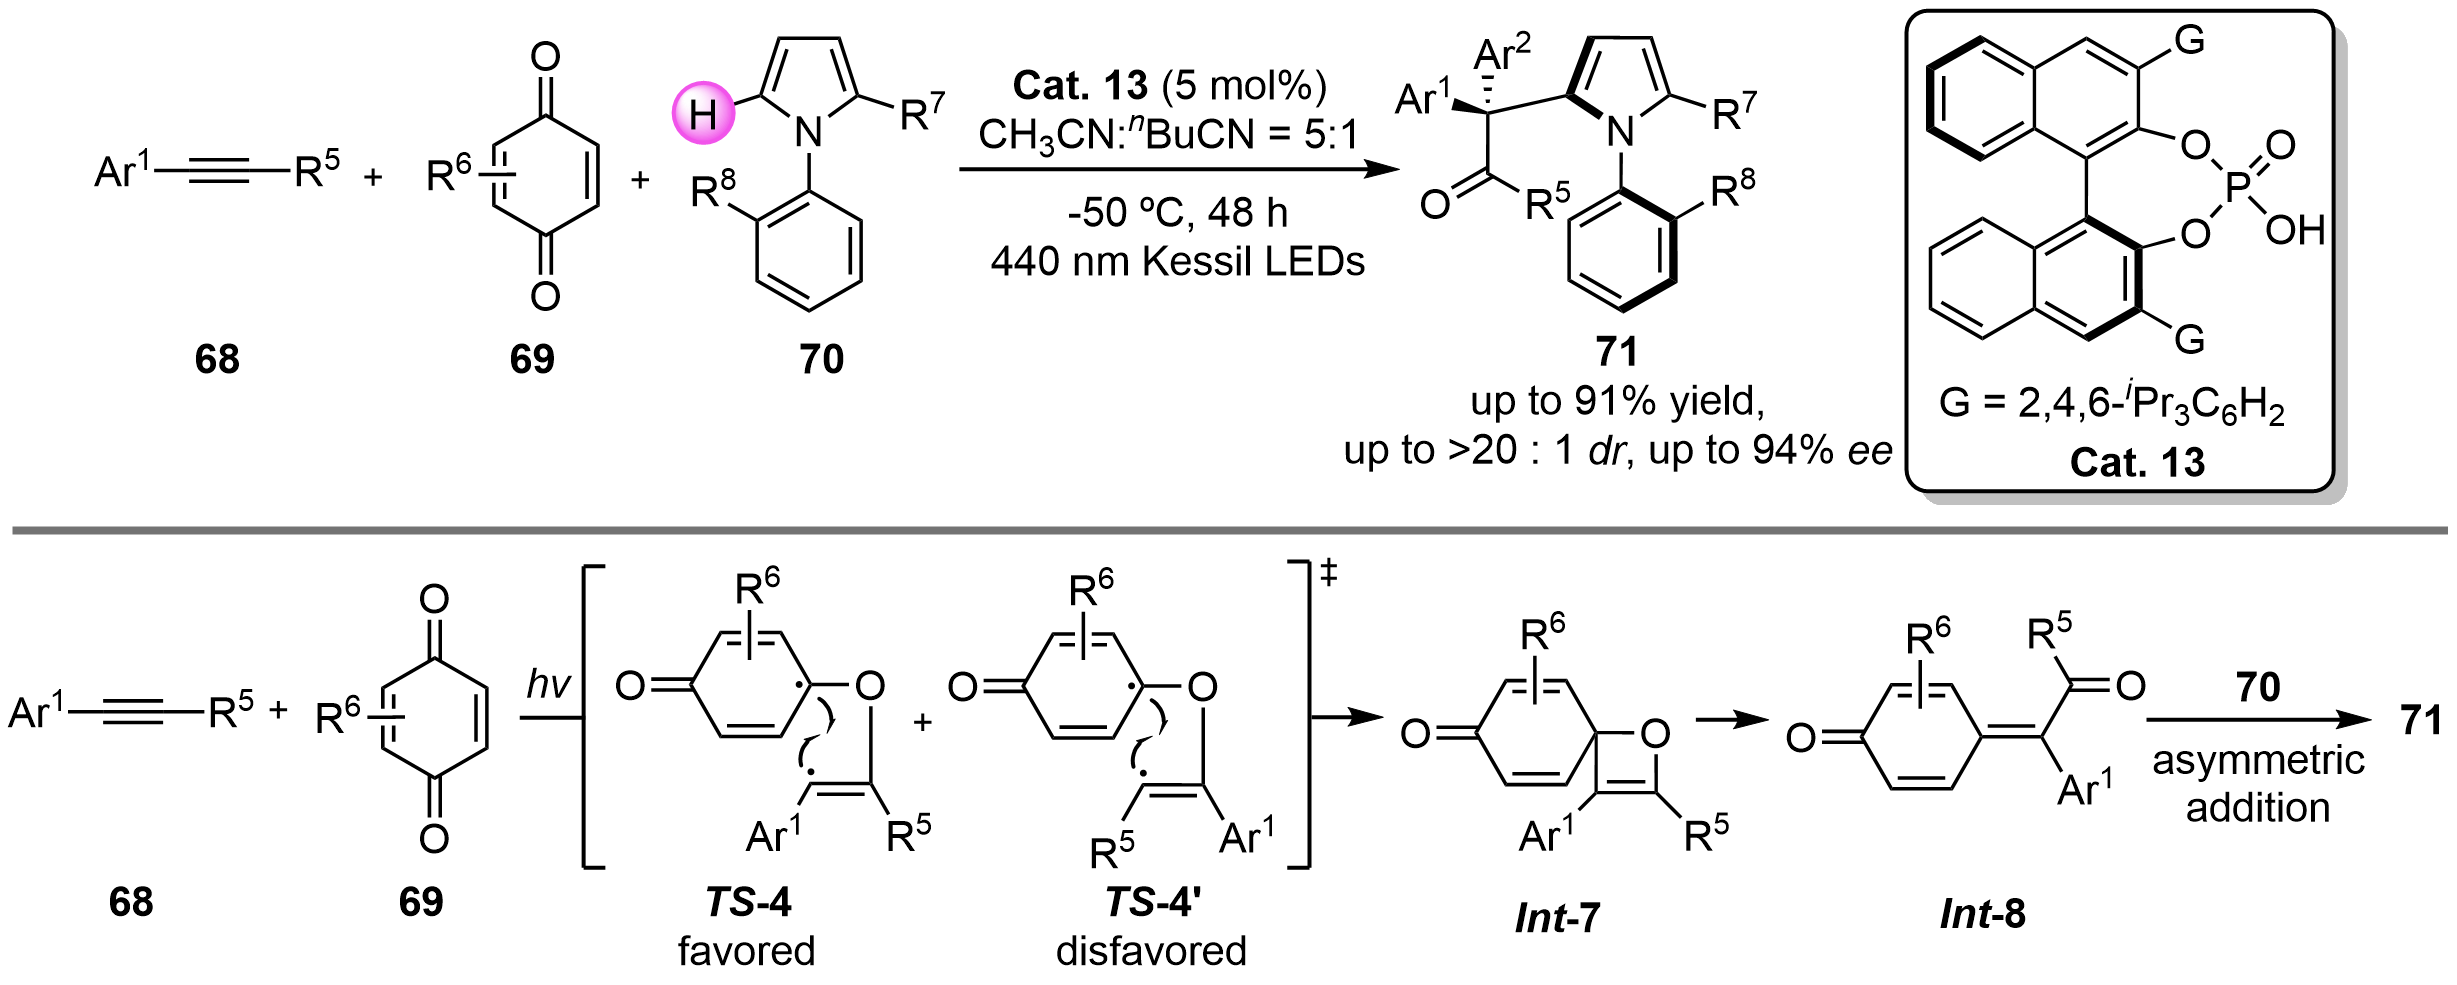
In 2023, Lu and co-workers conducted a chiral phosphoric acid-catalyzed asymmetric addition reaction to yield axially chiral N-arylpyrroles. The key intermediates Int-8 for the oxidation of alkynes by benzoquinone were formed through the cycloaddition/ring-opening reaction facilitated by light (Scheme 24)[83]. This exceptional regioselectivity was attributed to the fact that the benzylic radical possesses substantially greater stability than the alkyl radical in TS-4 and TS-4'. Therefore, the regioselective ring-opening cleavage preferentially occurs at the benzylic position to generate a thermodynamically more stable intermediate Int-8. Additionally, the steric hindrance between the bulky phenyl substituent and the smaller alkyl group plays a crucial role. The less sterically hindered alkyl-substituted site is preferentially attacked, which ultimately dictates the exclusive regioselectivity.
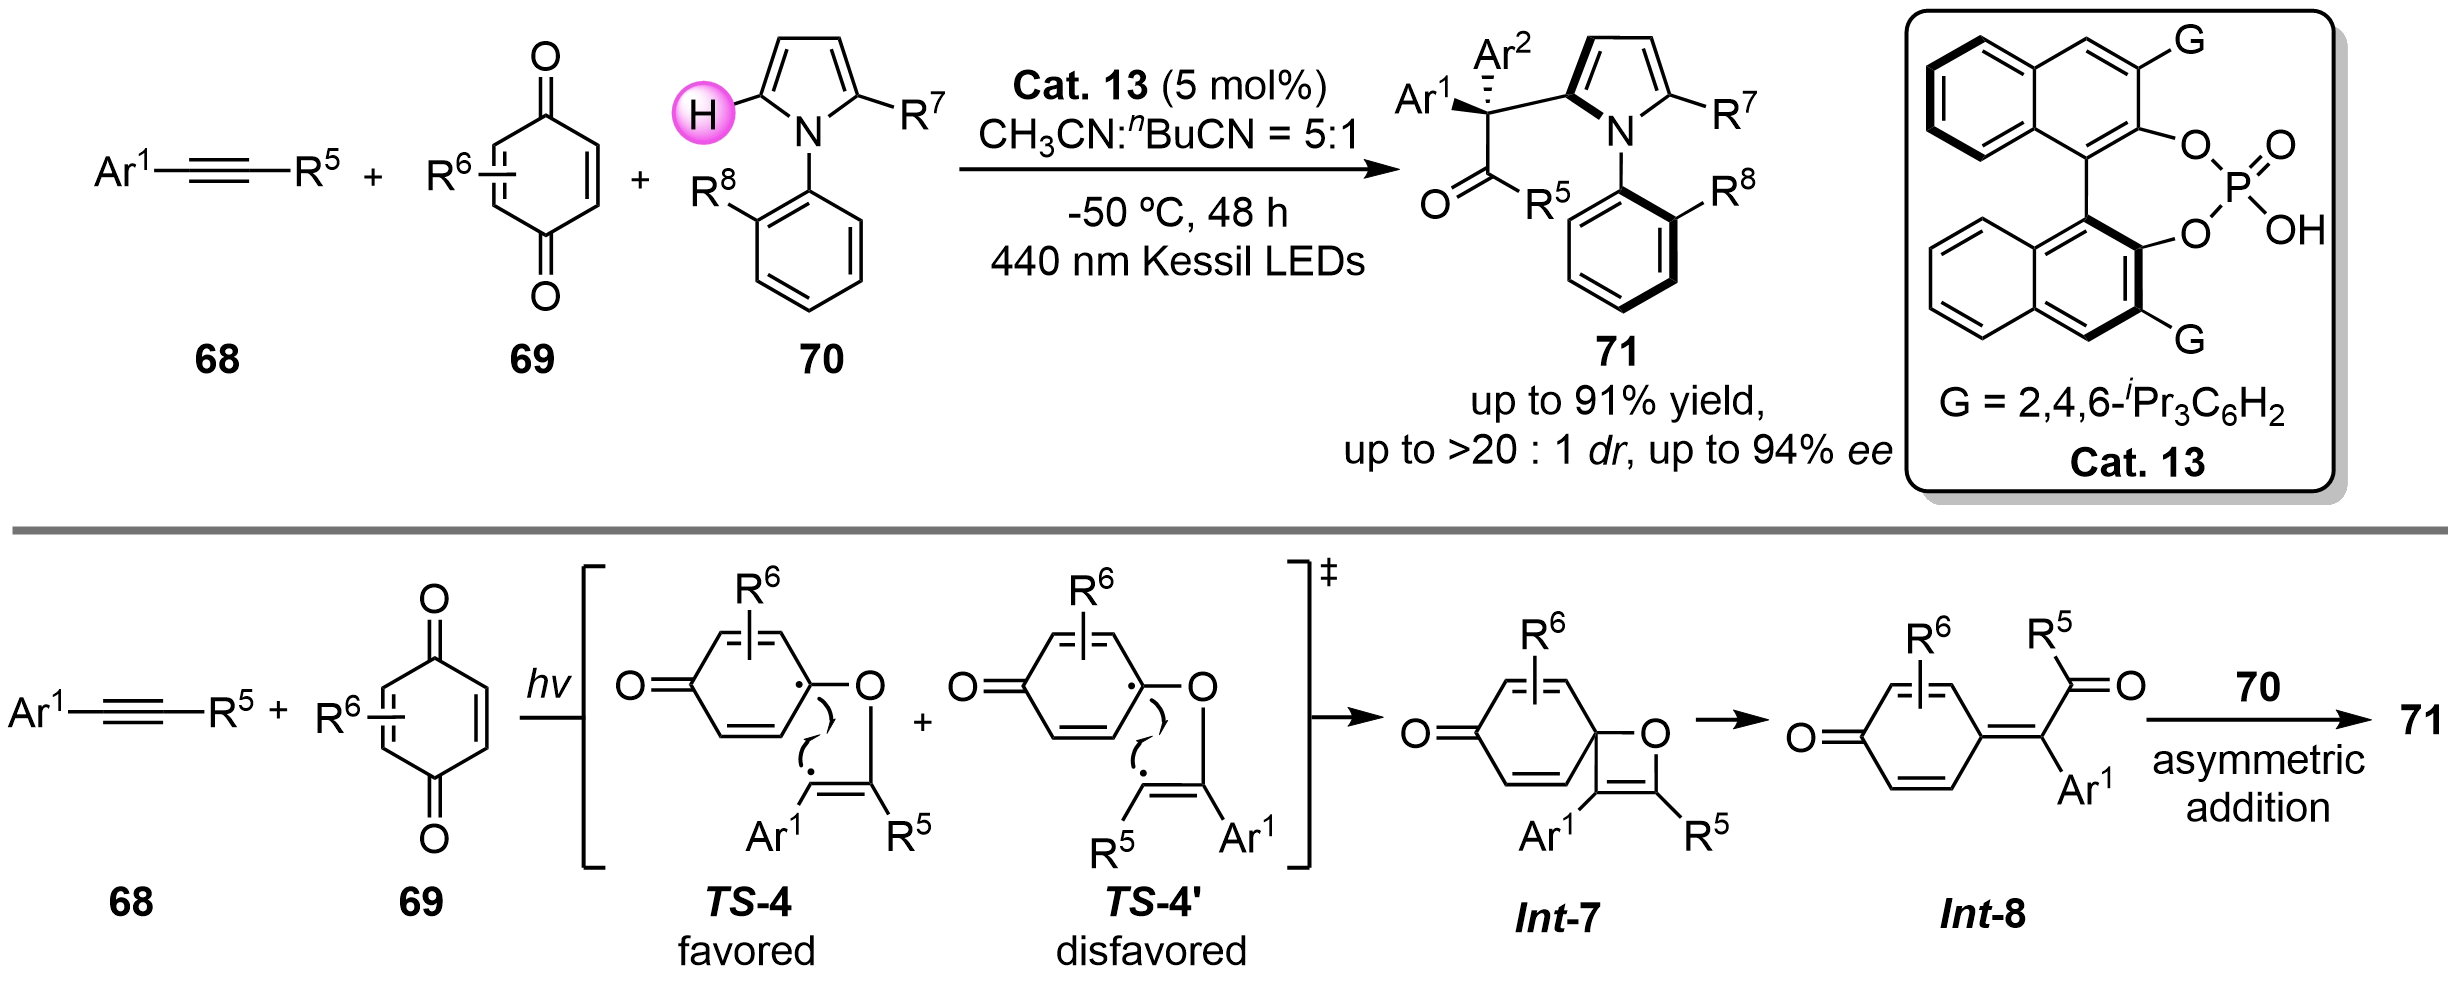
Scheme 24. Photocatalyzed one-pot simultaneous construction of central and axial chirality.
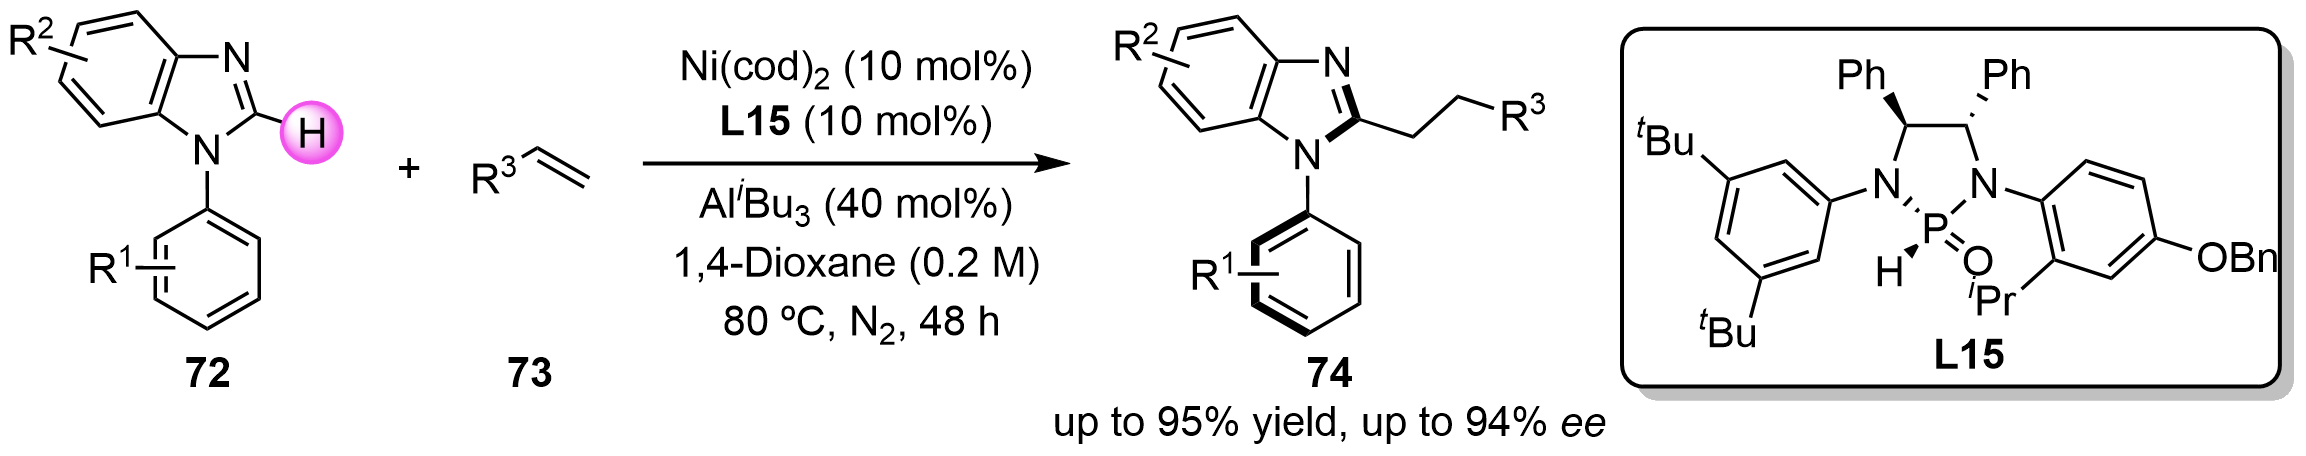
In 2024, Ackermann, Hong, and co-workers developed a distinctive a bimetallic Ni−Al catalytic system enabling C−H alkylation of 1-arylbenzimidazoles with terminal olefins, assisted by heteroatom-substituted secondary phosphine oxide (HASPO). In this reaction, AliBu3 serves as a Lewis acid to coordinate with the N-atom from benzimidazole substrate 72 to increase the C−H acidity in benzimidazoles, thereby promoting the nickel-mediated C−H functionalization reaction[84,85]. DFT calculations show that the LLHT mechanism is more favorable in energy than the classical oxidative addition and olefin insertion pathway, and the chiral ligand of HASPO plays a key role in the control of enantioselectivity (Scheme 25)[86].
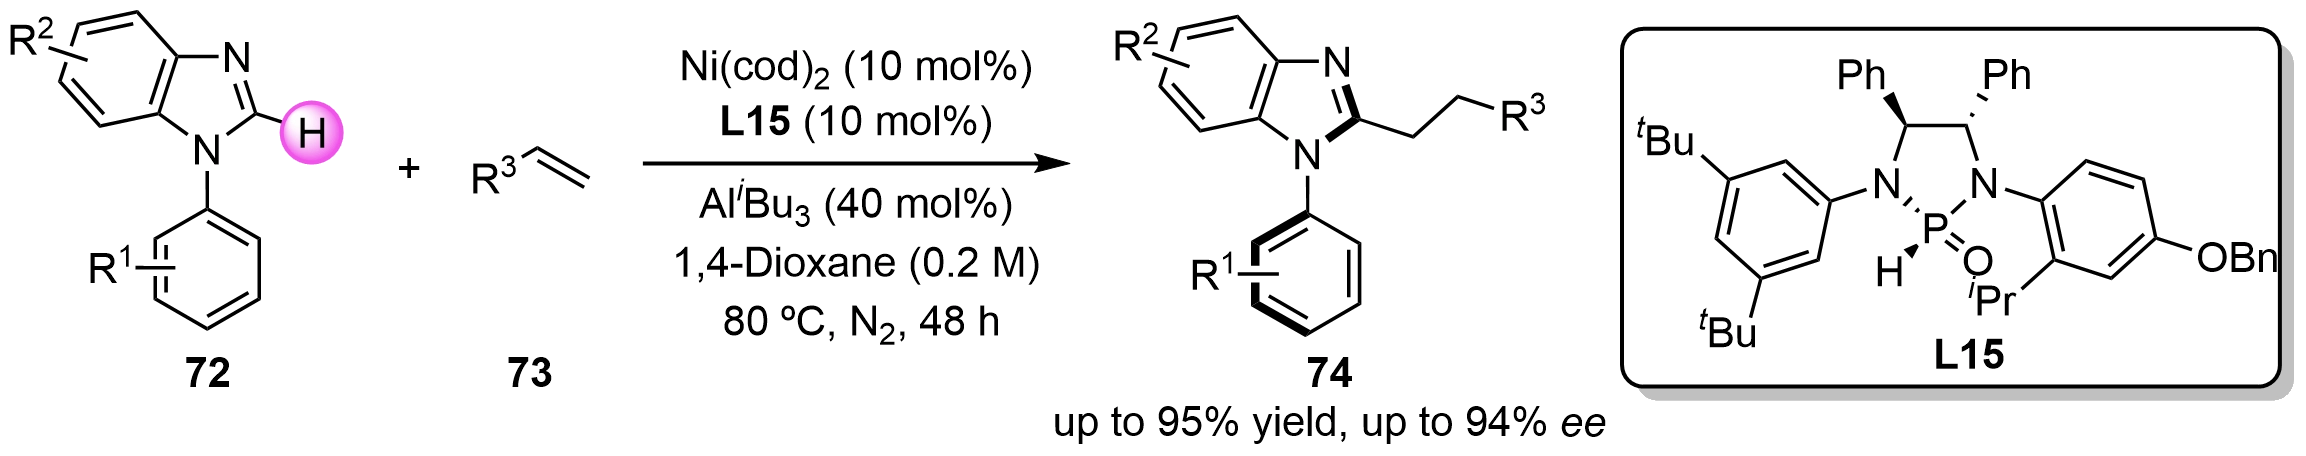
Scheme 25. Construction of C−N axial chirality by asymmetric C−H alkylation via LLHT pathway. LLHT: ligand-to-ligand hydrogen transfer.
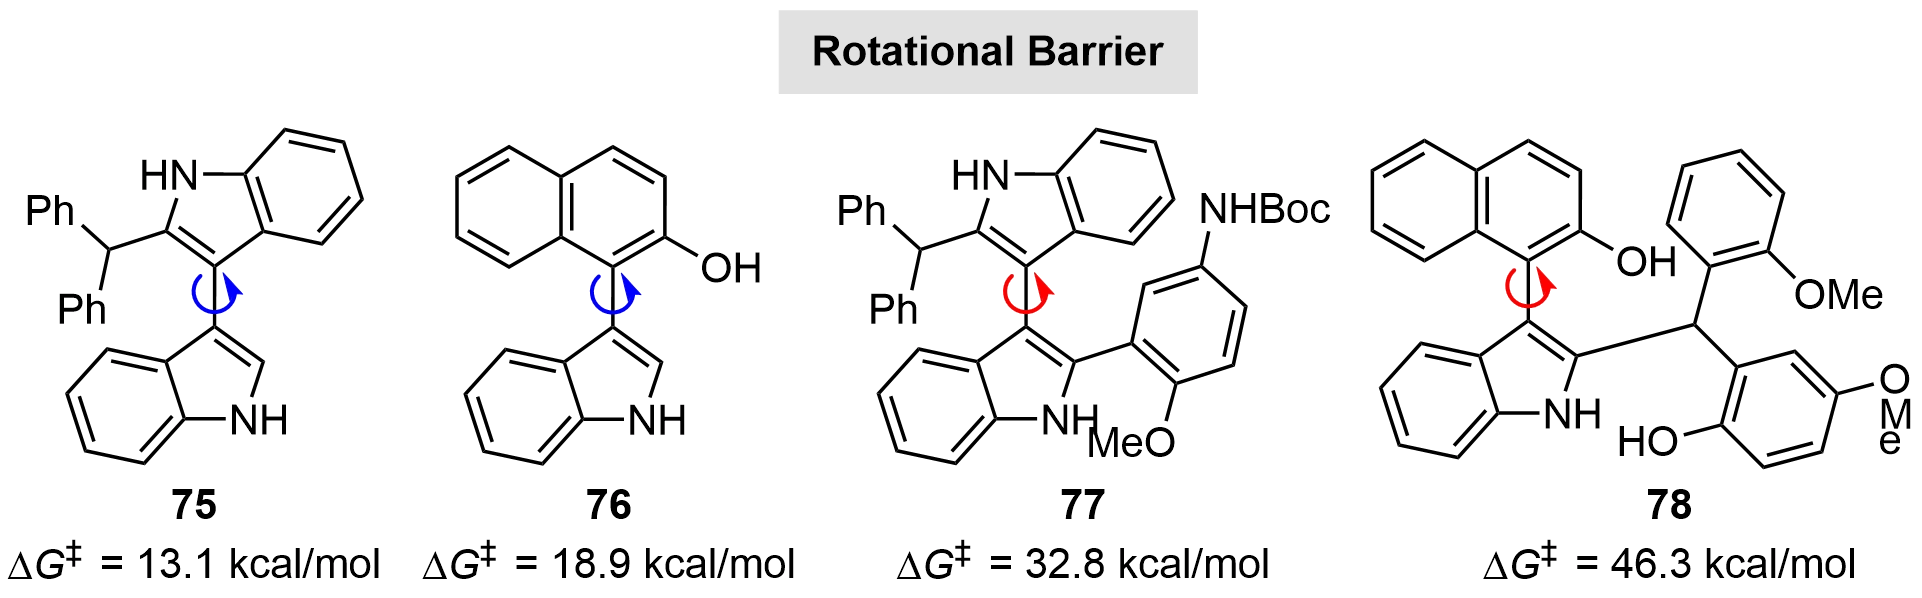
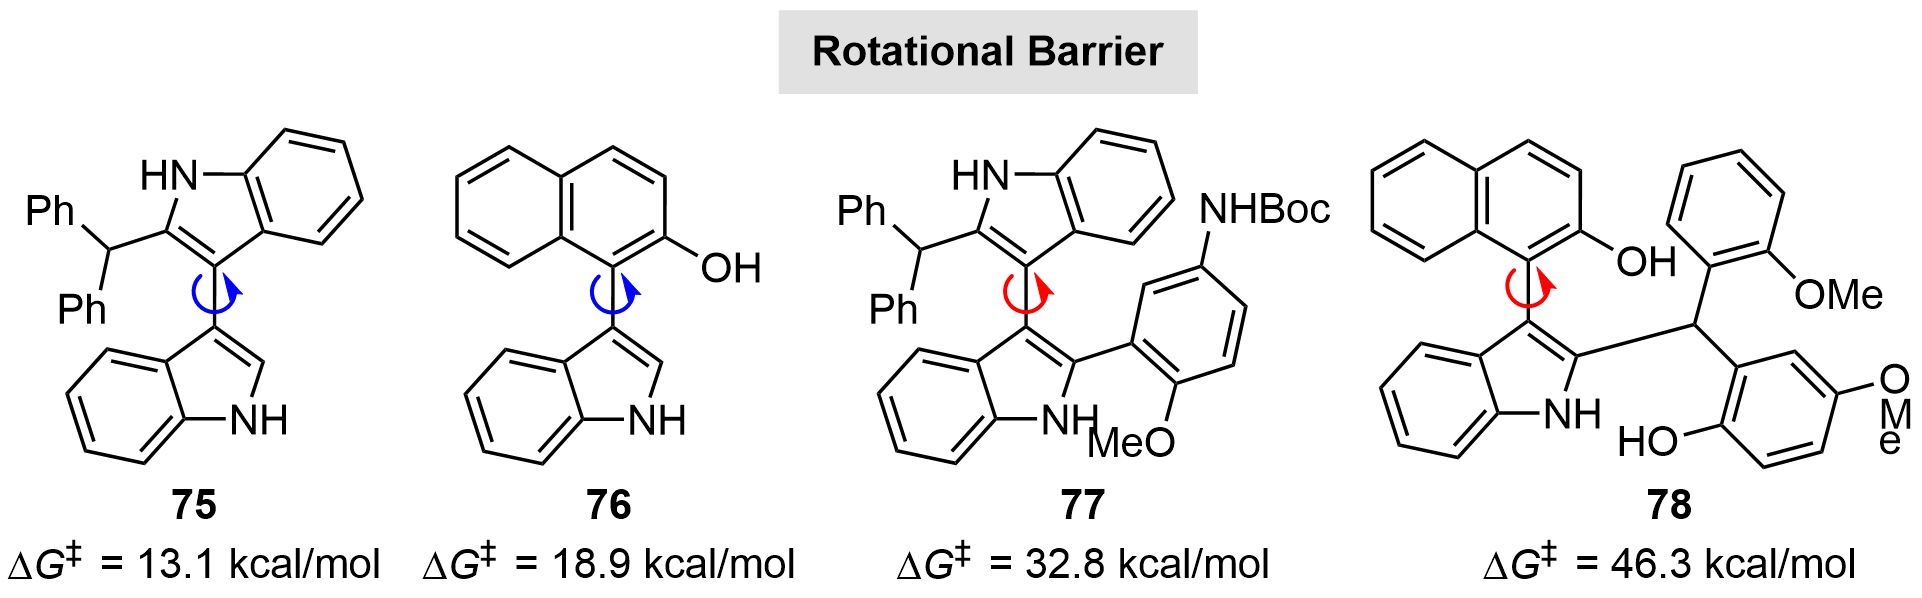
Aryl-indole or bisindole skeletons feature an electron-rich aromatic structure in the heteroaromatic system where the N−H moiety can be reserved for potentially forming a hydrogen bond. Bearing a smaller angle in the five-membered aromatic ring, the longer distance between the ortho-substituents on both sides of the axis leads to a decrease in the rotational energy barrier, thus resulting in the configurational instability of the axial chirality in aryl-indole or bisindole skeletons. For example, as illustrated in Scheme 26, when the C2-position of the indole ring remains unsubstituted within the bisindole or naphthyl-indole framework, the Shi group calculated the rotational barriers for 75 and 76 to be 13.1 kcal/mol[87] and 18.9 kcal/mol[88], respectively. These results confirm that these substrates readily undergo rapid racemization at room temperature, given that their rotational barriers fall below the 24 kcal/mol threshold required for the isolation of enantiomers[89]. Conversely, when bulky electrophiles are installed at the ortho-positions, the resulting severe steric congestion significantly restricts axial rotation. Consequently, for substrates 77 and 78, the rotational barriers markedly increase to over 30 kcal/mol, thereby endowing them with stable axial chirality[90].
In 2019, the Shi group developed an asymmetric nucleophilic substitution reaction, where tertiary alcohols were employed as the electrophiles and 3,3’-bisindoles were used as nucleophiles to conduct the attack at the C2 position of one indole side under the catalysis of chiral phosphoric acid Cat. 14. The DKR transformations show high efficiency and enantioselectivity with a broad scope for both the electrophiles and nucleophiles. The two large groups at the C2- and C2'-positions in products 81 became sterically hindered, so that the axis between the two indole rings was not able to rotate freely. In this way, the construction of axially chiral 3,3’-bisindole skeletons was successfully achieved. It is worth emphasizing that the chiral phosphoric acid is a bifunctional catalyst in this reaction, acting as a hydrogen bond donor and acceptor to activate both the nucleophilic and electrophilic units (Scheme 27)[87].
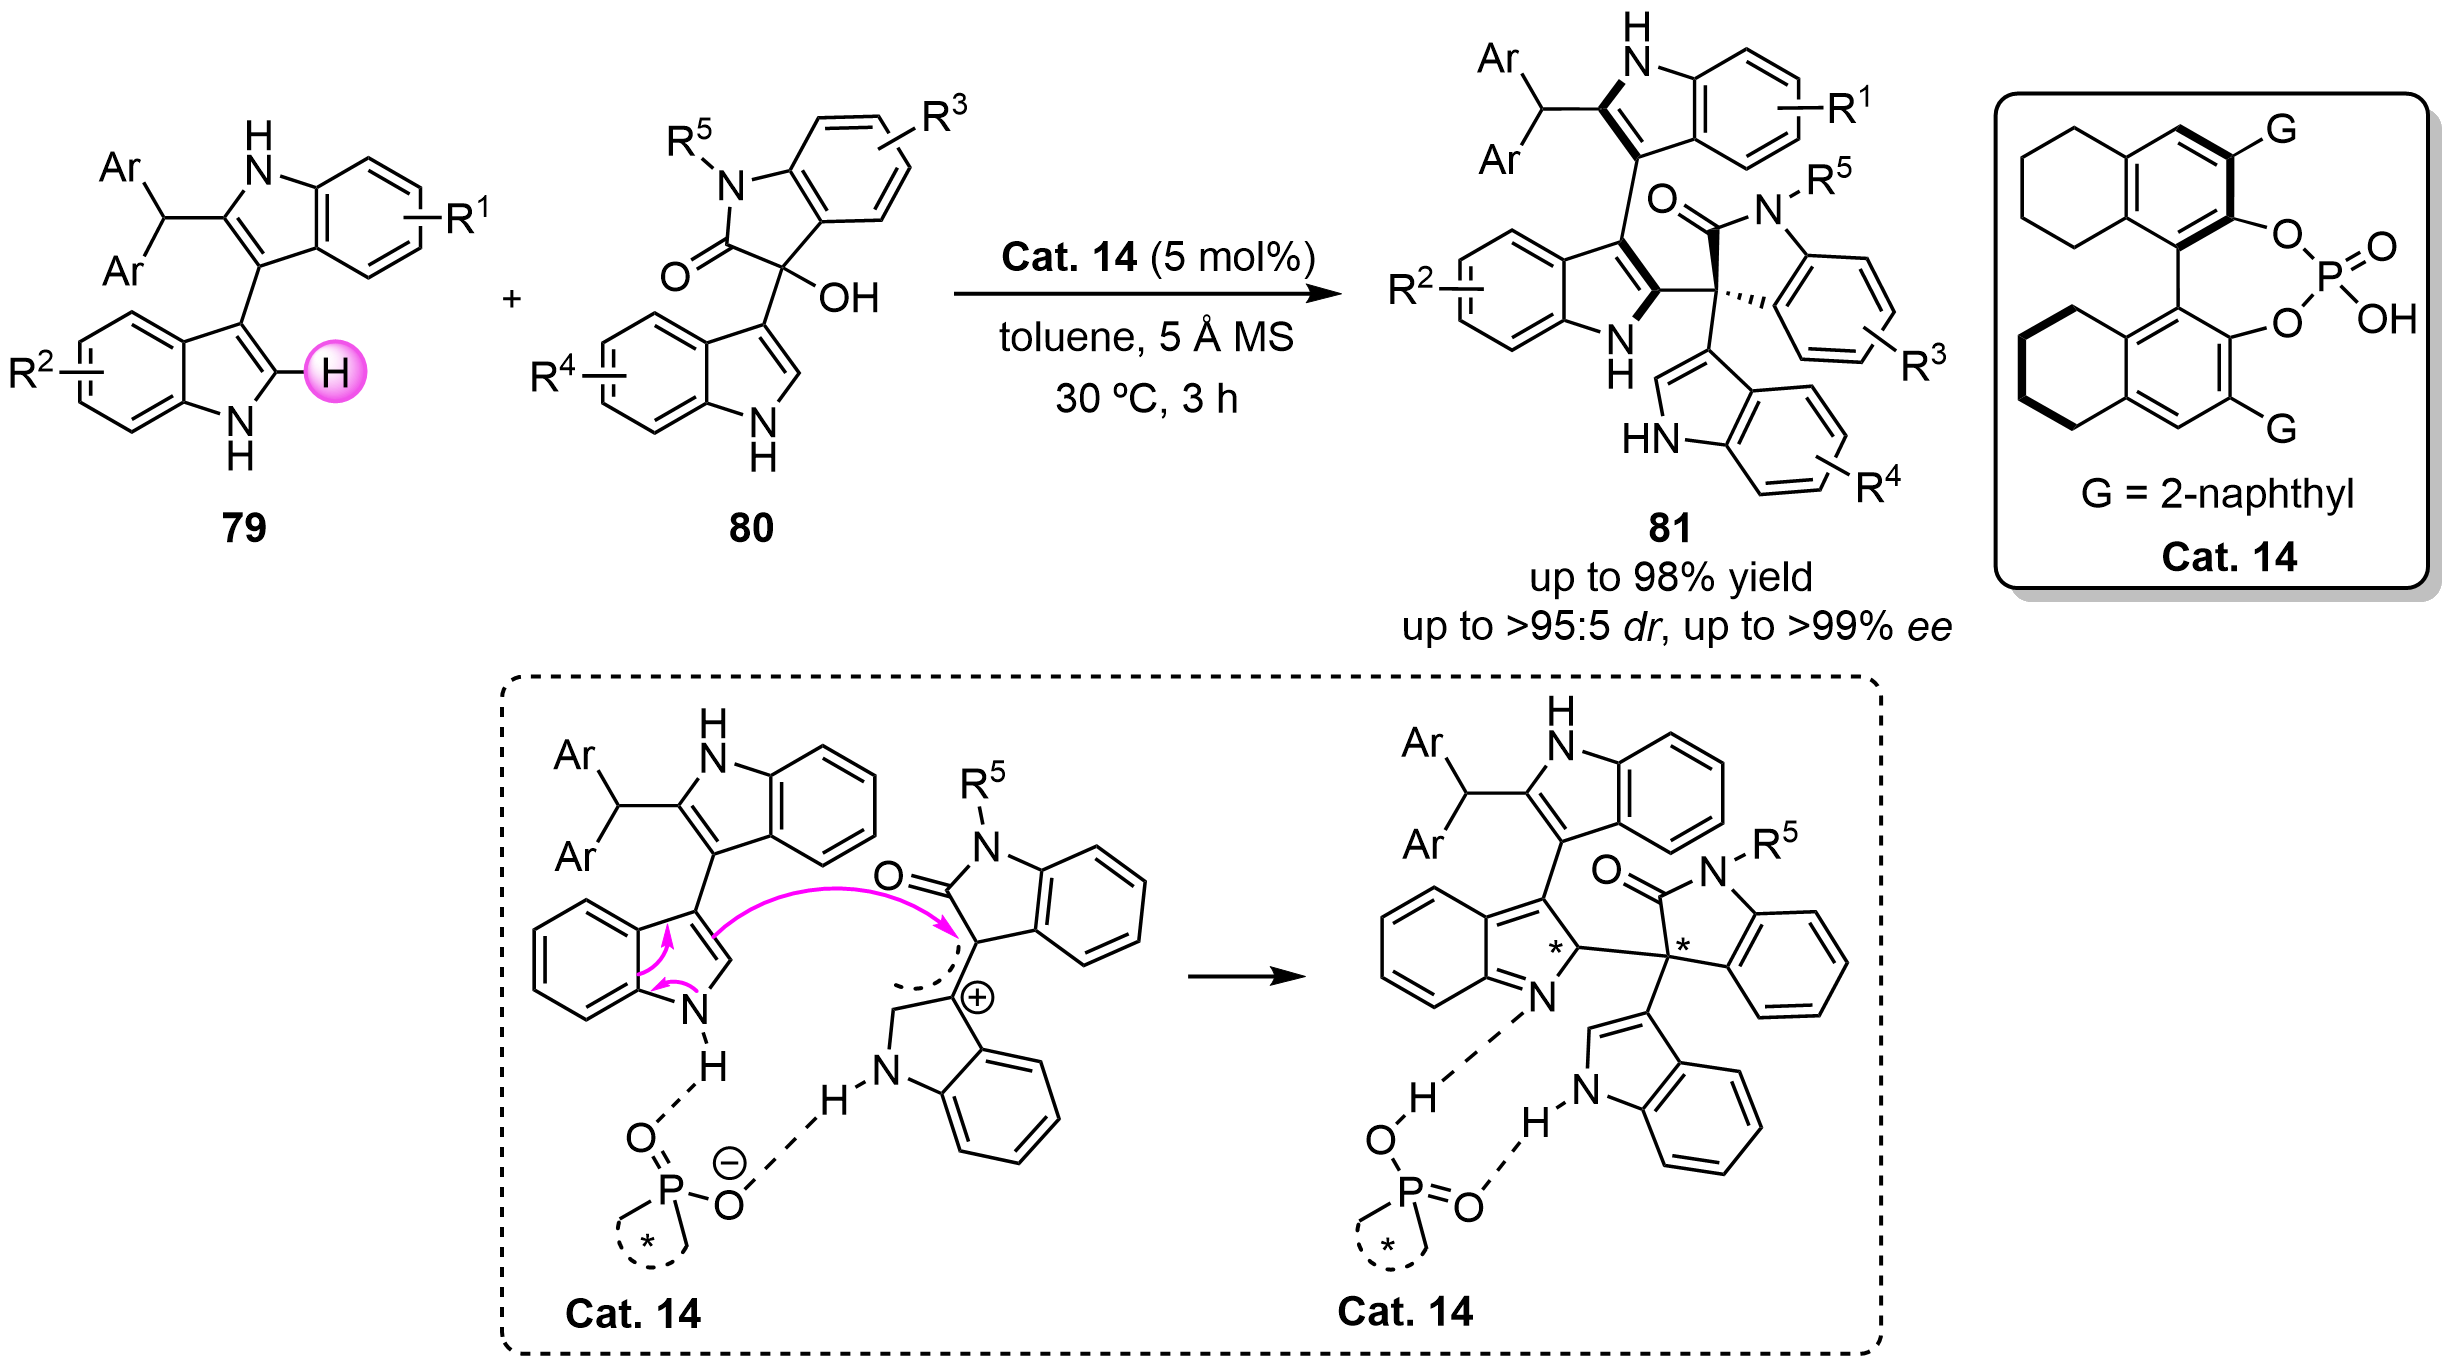
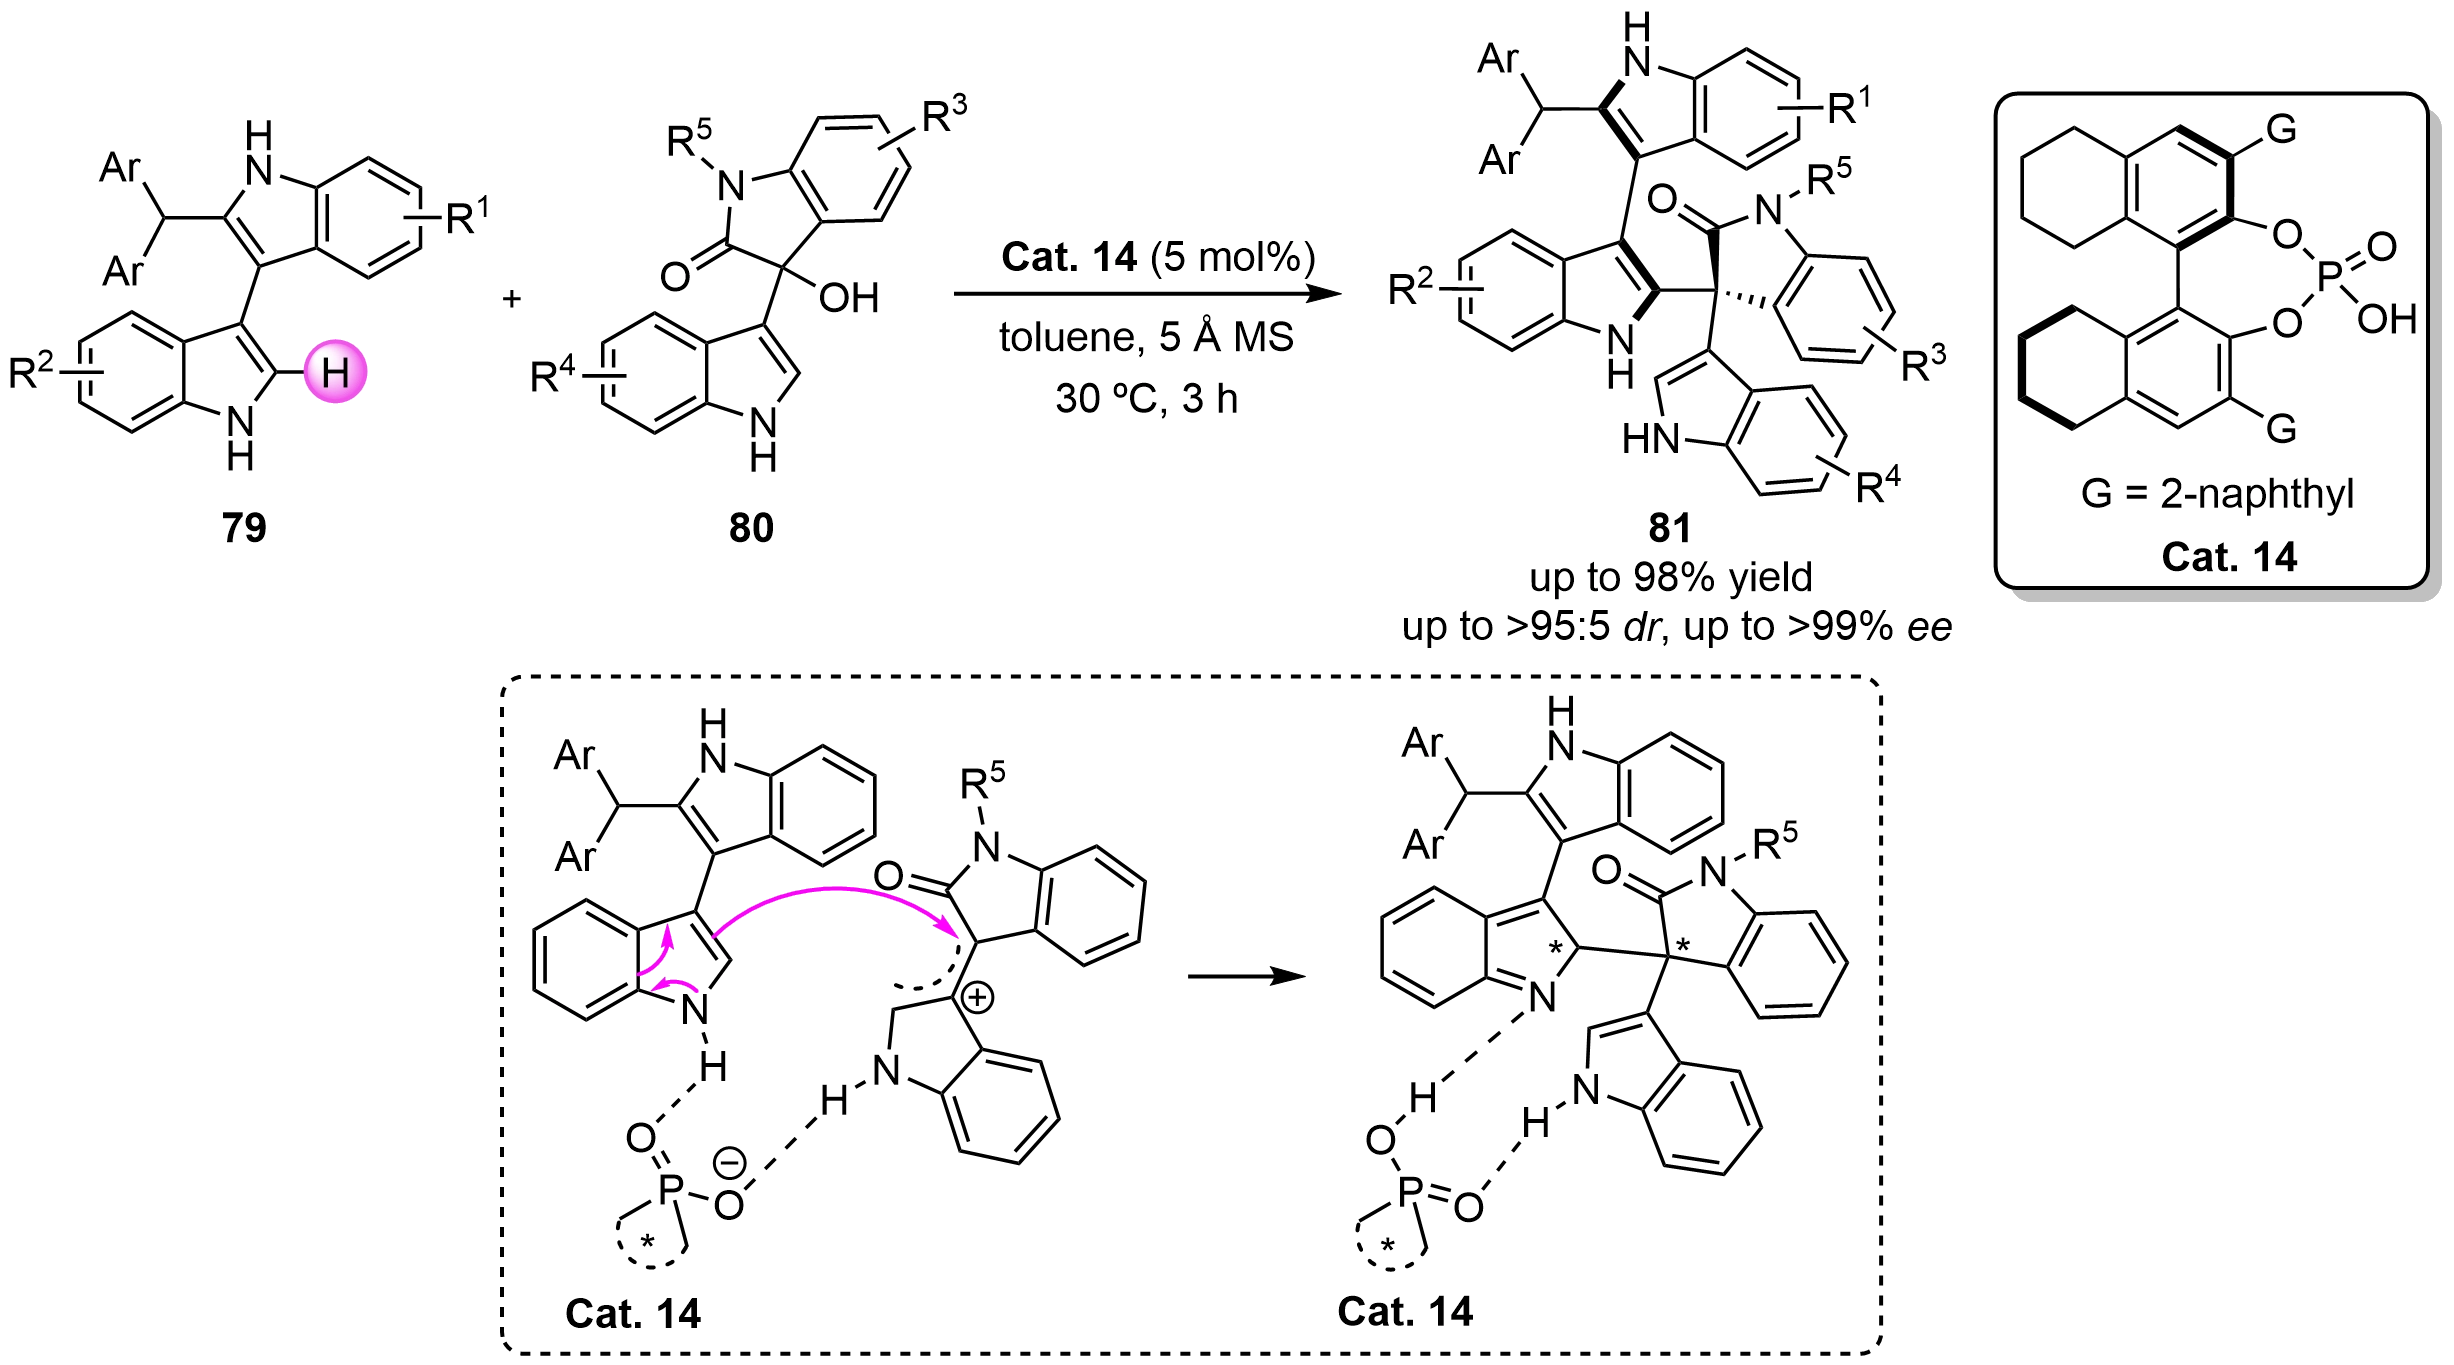
Scheme 27. DKR of bisindole skeletons enabled by chiral phosphoric acid catalyst. DKR: dynamic kinetic resolution.
Subsequently, the same research group employed azodicarboxylates, o-hydroxybenzyl alcohols[88], isatin-derived imines[91], propargylic alcohols[92], 3-alkynyl-3-hydroxyisoindolinones[93], and 2,3-indolyldimethanols[94] as electrophiles to develop the DKR of aryl-indole and bisindole skeletons through asymmetric substitutions (Scheme 28). In a similar way, other groups including the Fu[95], Kwon[96], Huang[97], and Li[98] also used chiral phosphoric acid to access axially chiral indole compounds. Quinine-derived chiral quaternary ammonium salts were also studied for the synthesis of axially chiral quinazolinones by the Jiang group[99]. The He group developed an amine-catalyzed asymmetric C−H alkylation of rotation-free 2-arylindoles with α,β-unsaturated aldehydes via the DKR process[100].
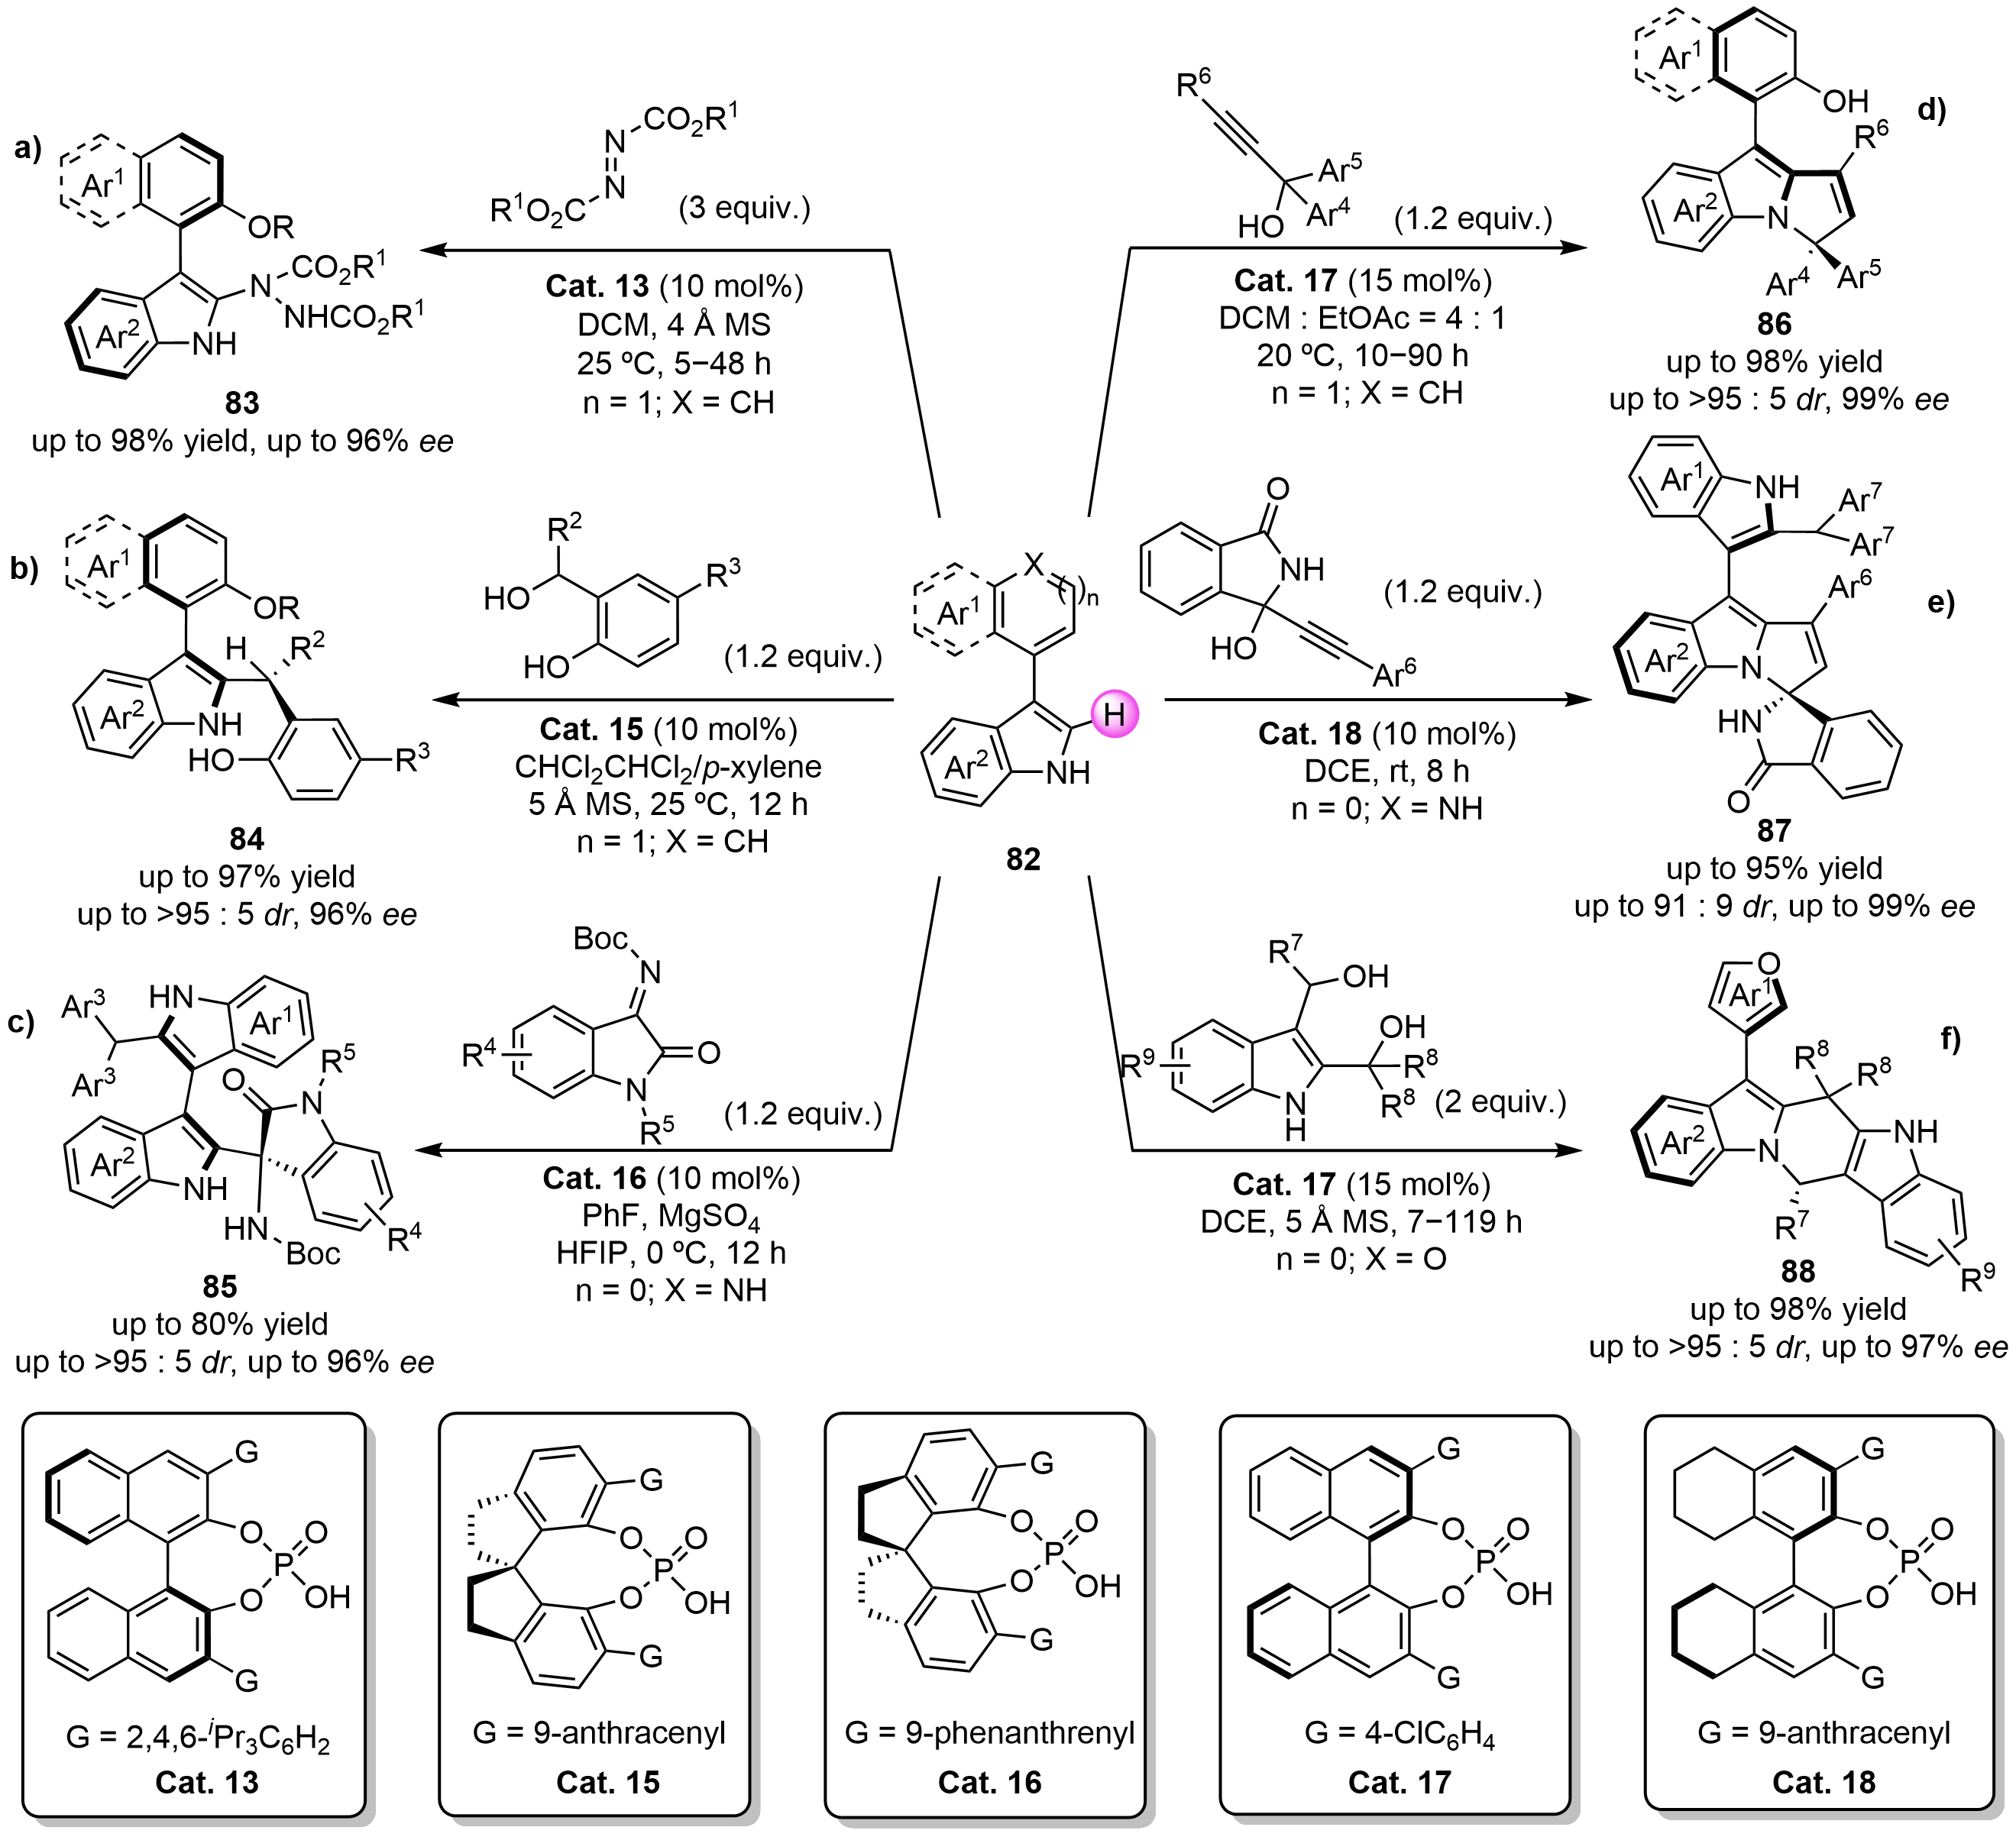
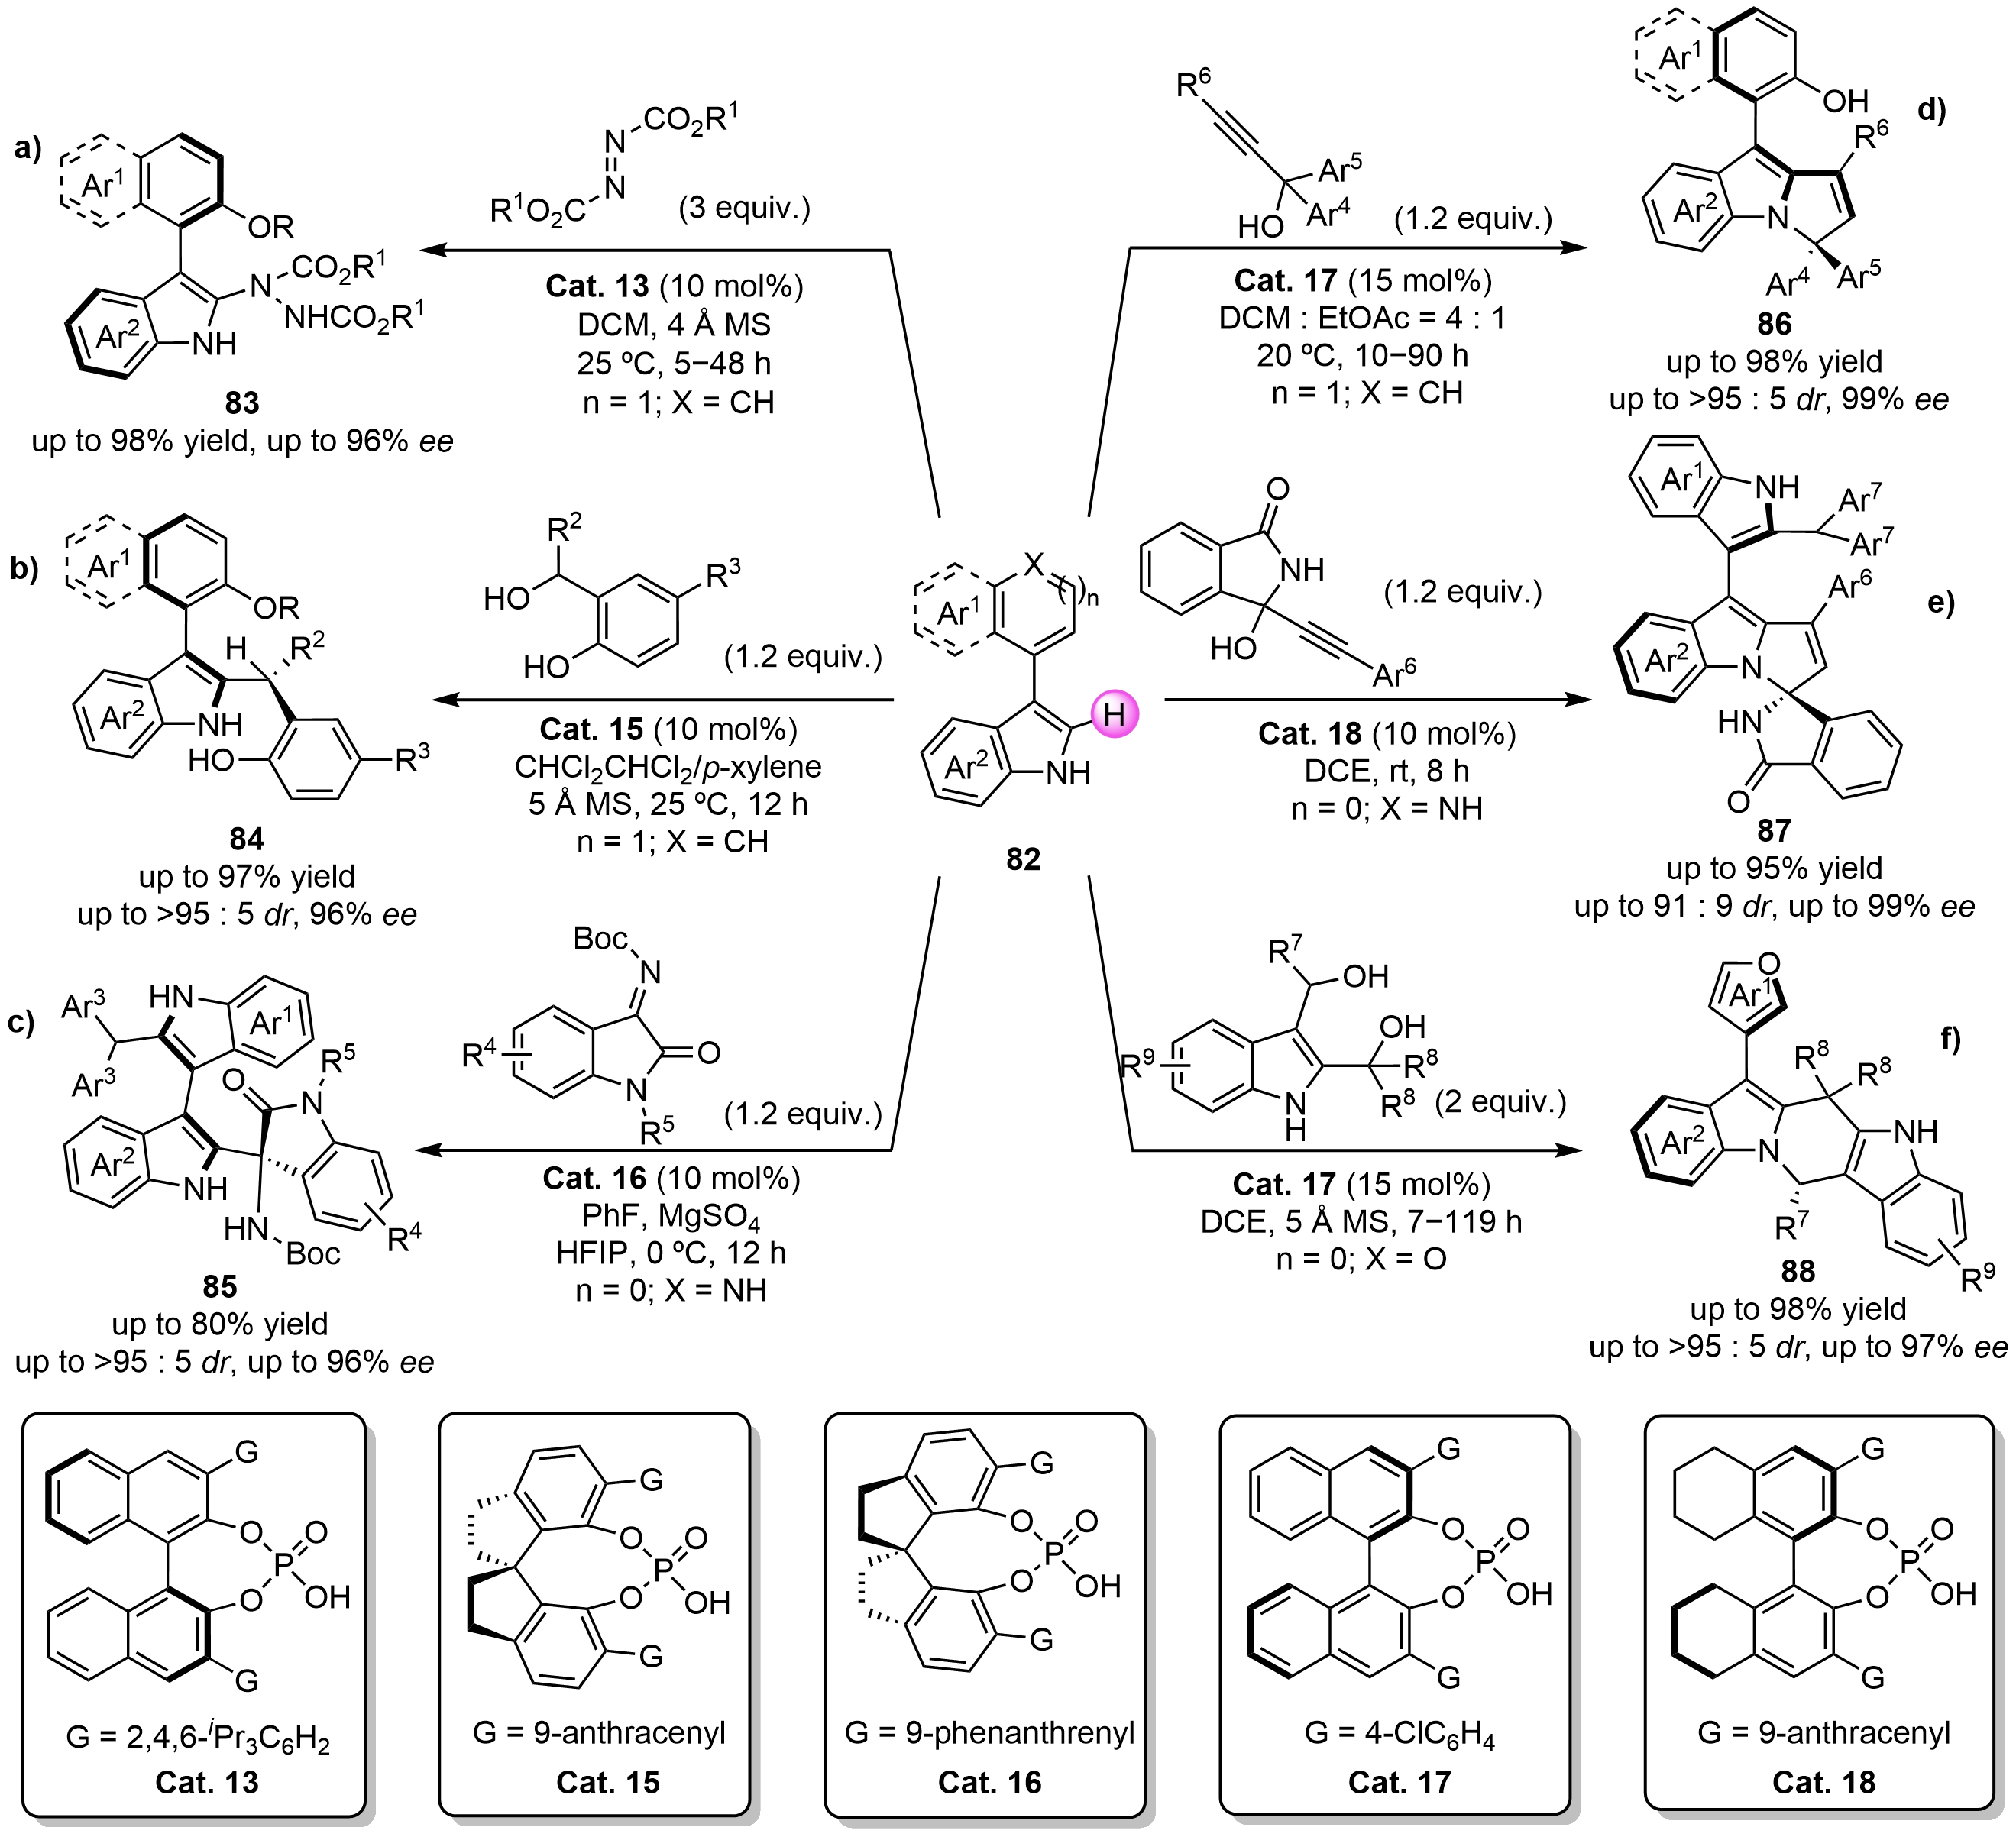
Scheme 28. DKR of indole-based biaryl skeletons enabled by chiral phosphoric acid catalyst. DKR: dynamic kinetic resolution.
3. Summary and Outlook
In summary, the DKR strategy via C−H bond activation has been demonstrated to be an important method for constructing such scaffolds with a theoretical yield of up to 100%. In this process, the starting material undergoes free rotation around the axis, while the product is conformationally locked by steric hindrance that restricts bond rotation.
This review summarizes the seminal contributions and recent advances in the construction of axially chiral atropisomeric biaryls via the DKR of configurationally labile biaryls utilizing C−H functionalization strategies. Three principal methodologies are systematically discussed: 1) Directed C−H functionalization: this approach employs coordinating directing groups, such as quinoline, isoquinoline, imine, oxime, aniline, pyridine, phosphorus center, P(=O), amides, esters, and thioethers to coordinate with metal catalysts and facilitate proximal C−H activation, which has proven to be effective for building chiral (hetero)aryl-aryl frameworks; 2) TDG strategy: by overcoming the limitations of fixed directing groups, the TDG strategy not only ensures precise stereocontrol but also broadens its scope from classical asymmetric catalysis to novel domains like electrosynthesis; 3) DG-free C−H functionalization: this approach has been successfully implemented in various reaction types, including electrophilic aromatic substitution, asymmetric addition, asymmetric substitution, and asymmetric coupling, thereby providing diverse synthetic pathways for the construction of chiral atropisomeric biaryls.
Despite the significant achievements outlined above, several critical challenges persist in the catalytic enantioselective synthesis of atropisomeric biaryls via C−H functionalization, which define key directions for future research: 1) Expanding directed C−H activation to remote sites: current strategies predominantly rely on coordination-guided functionalization at positions proximal to the directing group. To unlock novel chemical space, future efforts should focus on designing novel catalysts capable of recognizing and activating remote C−H bonds. This could be achieved by leveraging non-covalent secondary interactions, such as hydrogen bonding, to precisely steer the metal center toward distant sites. 2) Diversifying metal catalysts within the TDG paradigm: the development of TDG-assisted C−H functionalization has been overwhelmingly dominated by palladium catalysis. A highly promising direction for the field lies in exploring the compatibility of TDGs with alternative transition metals, thereby enabling a broader and more diverse spectrum of catalytic reaction modes. 3) Developing transition-metal-catalyzed C–H functionalization for DG-free strategies: successfully developed examples of DG-free C−H functionalization are primarily clustered around organocatalytic systems. In contrast, transition-metal-catalyzed C−H functionalization that operates without the aid of DGs remains relatively scarce in the field of DKR of configurationally labile biaryls via enantioselective C−H functionalization, representing an urgent need for the design of novel catalytic systems that achieve stereocontrol without substrate pre-functionalization.
Authors contribution
Zhang J, Huo X, Liu Y: Investigation, writing-original draft, writing-review & editing.
Hou XF, Zhu C: Supervision.
All authors have read and approved the final version of the manuscript.
Conflicts of interest
Not applicable.
Ethical approval
Not applicable.
Consent to participate
Not applicable.
Consent for publication
Not applicable.
Availability of data and materials
Not applicable.
Funding
We acknowledge the National Natural Science Foundation of China (Grant No. 22271054), the “1000-Youth Talents Plan”, Sinopec Seeding Program (ZC0607-0258). The project was also supported by Open Project of State Key Laboratory of Synergistic Chem-Bio Synthesis (sklscbs202510).
Copyright
© The Author(s) 2026.
References
-
1. Mei GJ, Koay WL, Guan CY, Lu Y. Atropisomers beyond the C–C axial chirality: Advances in catalytic asymmetric synthesis. Chem. 2022;8(7):1855-1893.[DOI]
-
3. Dong Y, Liu R, Wang W. Catalytic asymmetric Catellani-type reaction: A powerful tool for axial chirality construction. Green Synth Catal. 2020;1(2):83-85.[DOI]
-
4. Fu W, Tang W. Chiral monophosphorus ligands for asymmetric catalytic reactions. ACS Catal. 2016;6(8):4814-4858.[DOI]
-
6. Liang D, Xiao W, Lakhdar S, Chen J. Construction of axially chiral compounds via catalytic asymmetric radical reaction. Green Synth Catal. 2022;3(3):212-218.[DOI]
-
9. Woldegiorgis AG, Han Z, Lin X. Recent advances in chiral phosphoric acid catalyzed asymmetric organic reactions: An overview. J Mol Struct. 2024;1297:136919.[DOI]
-
11. Robinson DEJE, Bull SD. Kinetic resolution strategies using non-enzymatic catalysts. Tetrahedron Asymmetry. 2003;14(11):1407-1446.[DOI]
-
12. Lou D, Wu H, Zhu C. Design and construction of difunctionalized C−N atropisomers via lipase-catalyzed desymmetrization and kinetic resolution of N-naphthol-carbazoles. Chin Chem Lett. 2026;37(7):111943.[DOI]
-
13. Wu X, Zhao S, Che F, Wu X. Carbene-catalyzed enantioselective atherton-todd reaction for kinetic resolution of 1,1’-biaryl-2,2’-diols and derivatives. Org Lett. 2026;28(21):6785-6790.[DOI]
-
14. Xu Z, Zhou H, Chen X, Xu J. Kinetic resolution of unprotected BINOLs via Lewis base-catalyzed O-Boc protection. Org Lett. 2026;28(1):272-276.[DOI]
-
15. Zhang J, Zheng Z, Zhu C. Stereochemical editing: Catalytic racemization of secondary alcohols and amines. Chin Chem Lett. 2024;35(5):109160.[DOI]
-
16. Zhang J, Huo X, Liu Y, Zhu C. Dynamic kinetic resolution and dynamic kinetic asymmetric transformation of atropisomeric biaryls. Chem Catal. 2025;5(4):101329.[DOI]
-
22. Xiang SH, Ding WY, Wang YB, Tan B. Catalytic atroposelective synthesis. Nat Catal. 2024;7(5):483-498.[DOI]
-
23. Zhang HH, Li TZ, Liu SJ, Shi F. Catalytic asymmetric synthesis of atropisomers bearing multiple chiral elements: An emerging field. Angew Chem Int Ed. 2024;63(3):e202311053.[DOI]
-
24. Pedersen JR, Murto J, Korppi-Tommola J, Kuopio R, Pearson WB, Meisalo V. Activation parameters for inversion of 1-(1-naphthyl)-isoquinoline. I. Acta Chem Scand. 1972;26:929-936.[DOI]
-
25. Kakiuchi F, Le Gendre P, Yamada A, Ohtaki H, Murai S. Atropselective alkylation of biaryl compounds by means of transition metal-catalyzed C–H/olefin coupling. Tetrahedron Asymmetry. 2000;11(13):2647-2651.[DOI]
-
26. Albano G. Palladium-catalyzed cross-dehydrogenative coupling of (hetero)arenes. Org Chem Front. 2024;11(5):1495-1622.[DOI]
-
27. Jiang Q, Luo J, Zhao X. Enantioselective cross-dehydrogenative coupling enabled by organocatalysis. Green Chem. 2024;26(4):1846-1875.[DOI]
-
28. Patel DP, Kumar Singh S. Rose-Bengal-photocatalyzed cross-dehydrogenative coupling reactions under visible light. Eur J Org Chem. 2024;27(8):e202301185.[DOI]
-
34. Zhang WW, Wang Q, Zhang SZ, Zheng C, You SL. (SCp)rhodium-catalyzed asymmetric Satoh–Miura reaction for building-up axial chirality: Counteranion-directed switching of reaction pathways. Angew Chem Int Ed. 2023;62(3):e202214460.[DOI]
-
36. Gu Q, You SL, Zhou Q, Yin SY, Zheng DS, Zhang WW, et al. Enantioselective synthesis of axially chiral 1-arylisoquinolines by iridium(I)-catalyzed hydroarylation of alkynes. Synlett. 2023;34(12):1442-1446.[DOI]
-
37. Zheng DS, Zheng J, Wang Q, Gu Q, You SL. Rhodium(III)-catalyzed atroposelective C–H indolylization of 1-aryl isoquinolines with 3-indolylphenyliodonium salts. Chin J Chem. 2023;41(20):2684-2690.[DOI]
-
39. Zheng DS, Zhang WW, Gu Q, You SL. Rh(III)-catalyzed atroposelective C–H iodination of 1-aryl isoquinolines. ACS Catal. 2023;13(8):5127-5134.[DOI]
-
40. Zheng DS, Xie PP, Zhao F, Zheng C, Gu Q, You SL. Rh(III)-catalyzed atroposelective C–H selenylation of 1-aryl isoquinolines. ACS Catal. 2024;14(8):6009-6015.[DOI]
-
43. Luo J, Zhang T, Wang L, Liao G, Yao QJ, Wu YJ, et al. Enantioselective synthesis of biaryl atropisomers by Pd-catalyzed C–H olefination using chiral spiro phosphoric acid ligands. Angew Chem Int Ed. 2019;58(20):6708-6712.[DOI]
-
48. Wang L, Yuan WK, Wang ZK, Luo J, Zhou T, Shi BF. Synthesis of C–N axial chirality N-arylindoles via Pd(II)-catalyzed free amine-directed atroposelective C–H olefination. Chin J Chem. 2023;41(21):2788-2792.[DOI]
-
50. Liao G, Zhang T, Jin L, Wang BJ, Xu CK, Lan Y, et al. Experimental and computational studies on the directing ability of chalcogenoethers in palladium-catalyzed atroposelective C–H olefination and allylation. Angew Chem Int Ed. 2022;61(10):e202115221.[DOI]
-
52. Jiang BY, Zhou G, Jiang AL, Zhou T, Shi BF. Synthesis of axially chiral biaryl-2-carboxamides through Pd(ii)-catalyzed atroposelective C–H olefination. Org Chem Front. 2024;11(13):3710-3716.[DOI]
-
55. Wang YB, Tan B. Construction of axially chiral compounds via asymmetric organocatalysis. Acc Chem Res. 2018;51(2):534-547.[DOI]
-
56. Zhang HH, Wang CS, Li C, Mei GJ, Li Y, Shi F. Design and enantioselective construction of axially chiral naphthyl-indole skeletons. Angew Chem Int Ed. 2017;56(1):116-121.[DOI]
-
60. Li J, Long D, Liu H, Jiang J, Wang J. Expedient modular assembling of chiral η6-benzene ligands: Empowering ruthenium-catalyzed asymmetric C–H activation. J Am Chem Soc. 2026;148(1):1963-1976.[DOI]
-
61. Li Z, Wang M, Yang Y, Liang Y, Chen X, Zhao Y, et al. Atroposelective hydroarylation of biaryl phosphines directed by phosphorus centres. Nat Commun. 2023;14:8509.[DOI]
-
62. Jin S, Wang Y, Yan JX, Xu T, Ning M, Yuan Q, et al. P(═O)R2-directed asymmetric catalytic C–H olefination leading to C–N axially chiral targets. Org Lett. 2025;27(12):2838-2844.[DOI]
-
63. Qian PF, Wu YX, Hu JH, Chen JH, Zhou T, Yao QJ, et al. Atroposelective synthesis of pyridoindolones bearing two remote distinct C–N axes through cobalt-catalyzed enantioselective C–H activation. J Am Chem Soc. 2025;147(12):10791-10802.[DOI]
-
66. Liao G, Li B, Chen HM, Yao QJ, Xia YN, Luo J, et al. Pd-catalyzed atroposelective C–H allylation through β-O elimination: Diverse synthesis of axially chiral biaryls. Angew Chem Int Ed. 2018;57(52):17151-17155.[DOI]
-
67. Chen HM, Zhang S, Liao G, Yao QJ, Xu XT, Zhang K, et al. Pd-catalyzed atroposelective C–H allylation and alkenylation: Access to enantioenriched atropisomers featuring pentatomic heteroaromatics. Organometallics. 2019;38(20):4022-4028.[DOI]
-
69. Chen HM, Liao G, Xu CK, Yao QJ, Zhang S, Shi BF. Merging C–H and C–C activation in Pd(II)-catalyzed enantioselective synthesis of axially chiral biaryls. CCS Chem. 2021;3(12):455-465.[DOI]
-
70. Arjun V, Kaushik K, Jeganmohan M. Chiral transient directing group enabled palladium-catalyzed atroposelective C–H vinylation of biaryl aldehydes with vinyl silane. Org Lett. 2025;27(31):8452-8458.[DOI]
-
71. Zhang S, Yao QJ, Liao G, Li X, Li H, Chen HM, et al. Enantioselective synthesis of atropisomers featuring pentatomic heteroaromatics by Pd-catalyzed C–H alkynylation. ACS Catal. 2019;9(3):1956-1961.[DOI]
-
79. Miyaji R, Asano K, Matsubara S. Induction of axial chirality in 8-arylquinolines through halogenation reactions using bifunctional organocatalysts. Chemistry A European J. 2017;23(42):9996-10000.[DOI]
-
81. He C, Hou M, Zhu Z, Gu Z. Enantioselective synthesis of indole-based biaryl atropisomers via palladium-catalyzed dynamic kinetic intramolecular C–H cyclization. ACS Catal. 2017;7(8):5316-5320.[DOI]
-
88. Jiang F, Chen KW, Wu P, Zhang YC, Jiao Y, Shi F. A strategy for synthesizing axially chiral naphthyl-indoles: Catalytic asymmetric addition reactions of racemic substrates. Angew Chem Int Ed. 2019;58(42):15104-15110.[DOI]
-
91. Sheng FT, Li ZM, Zhang YZ, Sun LX, Zhang YC, Tan W, et al. Atroposelective synthesis of 3,3’-bisindoles bearing axial and central chirality: Using isatin-derived imines as electrophiles. Chin J Chem. 2020;38(6):583-589.[DOI]
-
92. Wu P, Yu L, Gao CH, Cheng Q, Deng S, Jiao Y, et al. Design and synthesis of axially chiral aryl-pyrroloindoles via the strategy of organocatalytic asymmetric (2 + 3) cyclization. Fundam Res. 2023;3(2):237-248.[DOI]
-
94. Wang JY, Gao CH, Ma C, Wu XY, Ni SF, Tan W, et al. Design and catalytic asymmetric synthesis of furan-indole compounds bearing both axial and central chirality. Angew Chem Int Ed. 2024;63(8):e202316454.[DOI]
-
96. Kim A, Kim A, Park S, Kim S, Jo H, Ok KM, et al. Catalytic and enantioselective control of the C–N stereogenic axis via the pictet–spengler reaction. Angew Chem Int Ed. 2021;60(22):12279-12283.[DOI]
-
97. Li C, Zuo WF, Zhou J, Zhou WJ, Wang M, Li X, et al. Catalytic asymmetric synthesis of 3, 4’-indole–pyrazole derivatives featuring axially chiral bis-pentatomic heteroaryls. Org Chem Front. 2022;9(7):1808-1813.[DOI]
-
98. Xia Y, Liu M, Qian C, Li P, Dong M, Li W. Asymmetric organocatalytic (3 + 2) annulation of propargylic alcohols with indolylnaphthalenols: Synergistic construction of axial and central chirality. Org Chem Front. 2022;10(1):30-34.[DOI]
-
100. Deng J, Li W, Zhou C, Li Z, Zhou H, Yan J, et al. A catalytic atroposelective Friedel–Crafts alkylation to access axially chiral C2-arylindoles via dynamic kinetic resolutions. Org Chem Front. 2025;12(2):584-590.[DOI]
Copyright
© The Author(s) 2027. This is an Open Access article licensed under a Creative Commons Attribution 4.0 International License (https://creativecommons.org/licenses/by/4.0/), which permits unrestricted use, sharing, adaptation, distribution and reproduction in any medium or format, for any purpose, even commercially, as long as you give appropriate credit to the original author(s) and the source, provide a link to the Creative Commons license, and indicate if changes were made.
Publisher’s Note
Science Exploration remains a neutral stance on jurisdictional claims in published
maps
and institutional affiliations. The views expressed in this article are solely those
of
the author(s) and do not reflect the opinions of the Editors or the publisher.
Share And Cite


Science Exploration Style
Zhang J, Huo X, Liu Y, Hou XF, Zhu C. Dynamic kinetic resolution (DKR) of configurationally labile biaryls via enantioselective C−H functionalization. Chiral Chem. 2027;3:202627. https://doi.org/10.70401/cc.2026.0036
Tips
Copy completed.
Submit a Manuscript
Author Instructions
Cite this Article
Article Metrics
0
View
0
Download
Cited
Article Updates

 Get citation
Science Exploration Style

Zhang J, Huo X, Liu Y, Hou XF, Zhu C. Dynamic kinetic resolution (DKR) of configurationally labile biaryls via enantioselective C−H functionalization. Chiral Chem. 2027;3:202627. https://doi.org/10.70401/cc.2026.0036
copy
Share Link
copy