Catalytic asymmetric synthesis of chiral carboxylic acids and heterocycles with CO2

Deyong Yang
1
,
Han Zhou
1
,
Lele Wang
1
,
Huimin Wang
1
,
Mengmeng Jiang
1
,
Yixiao Zhang
1
,
Dingying Chen
1
,
Xianglin Pan
2
,
Jian Zhou
1

,
Feng Zhou
1,*
*Correspondence to:
Feng Zhou, State Key Laboratory of Petroleum Molecular & Process Engineering, Shanghai Key Laboratory of Green Chemistry and Chemical Processes, Shanghai Engineering Research Center of Molecular Therapeutics and New Drug Development, School of Chemistry and Molecular Engineering, East China Normal University, Shanghai 200062, China.
E-mail: fzhou@chem.ecnu.edu.cn
Chiral Chem. 2027;3:202633. 10.70401/cc.2026.0038
Received: July 18, 2026Accepted: September 07, 2026Published: September 07, 2026
Abstract
The development of asymmetric catalytic methods using CO2 as a C1 synthon is an important strategy for constructing chiral carboxylic acids and heterocyclic compounds. In recent years, through the design of innovative catalytic systems, efficient and highly selective syntheses of chiral carboxylic acids, spanning central to axial chirality, have been achieved from diverse substrates, including alkenes, alkynes, and organic electrophiles. Concurrently, significant progress has also been made in the synthesis of chiral heterocycles, particularly 2-oxazolidinones and cyclic carbonates. This review systematically summarizes key advances in this field from 2021 to mid-2026. Organized by target product structure, the discussion first addresses chiral carboxylic acids, categorized by substrate type (carbon-carbon unsaturated bonds and organic electrophiles). Key strategies, including carbo-carboxylation, boracarboxylation, hydrocarboxylation, dicarboxylation, and electroreductive carboxylation, are elaborated in detail, along with their stereoselectivity control mechanisms. The discussion then covers chiral heterocycles, summarizing the catalytic asymmetric construction of chiral 2-oxazolidinones and chiral cyclic carbonates. Finally, the major challenges and future directions in this field are discussed.
Graphical Abstract
Keywords
Carbon dioxide fixation, C1 synthons, asymmetric catalysis, enantioselective carboxylation, chiral carboxylic acids, chiral heterocycles
1. Introduction
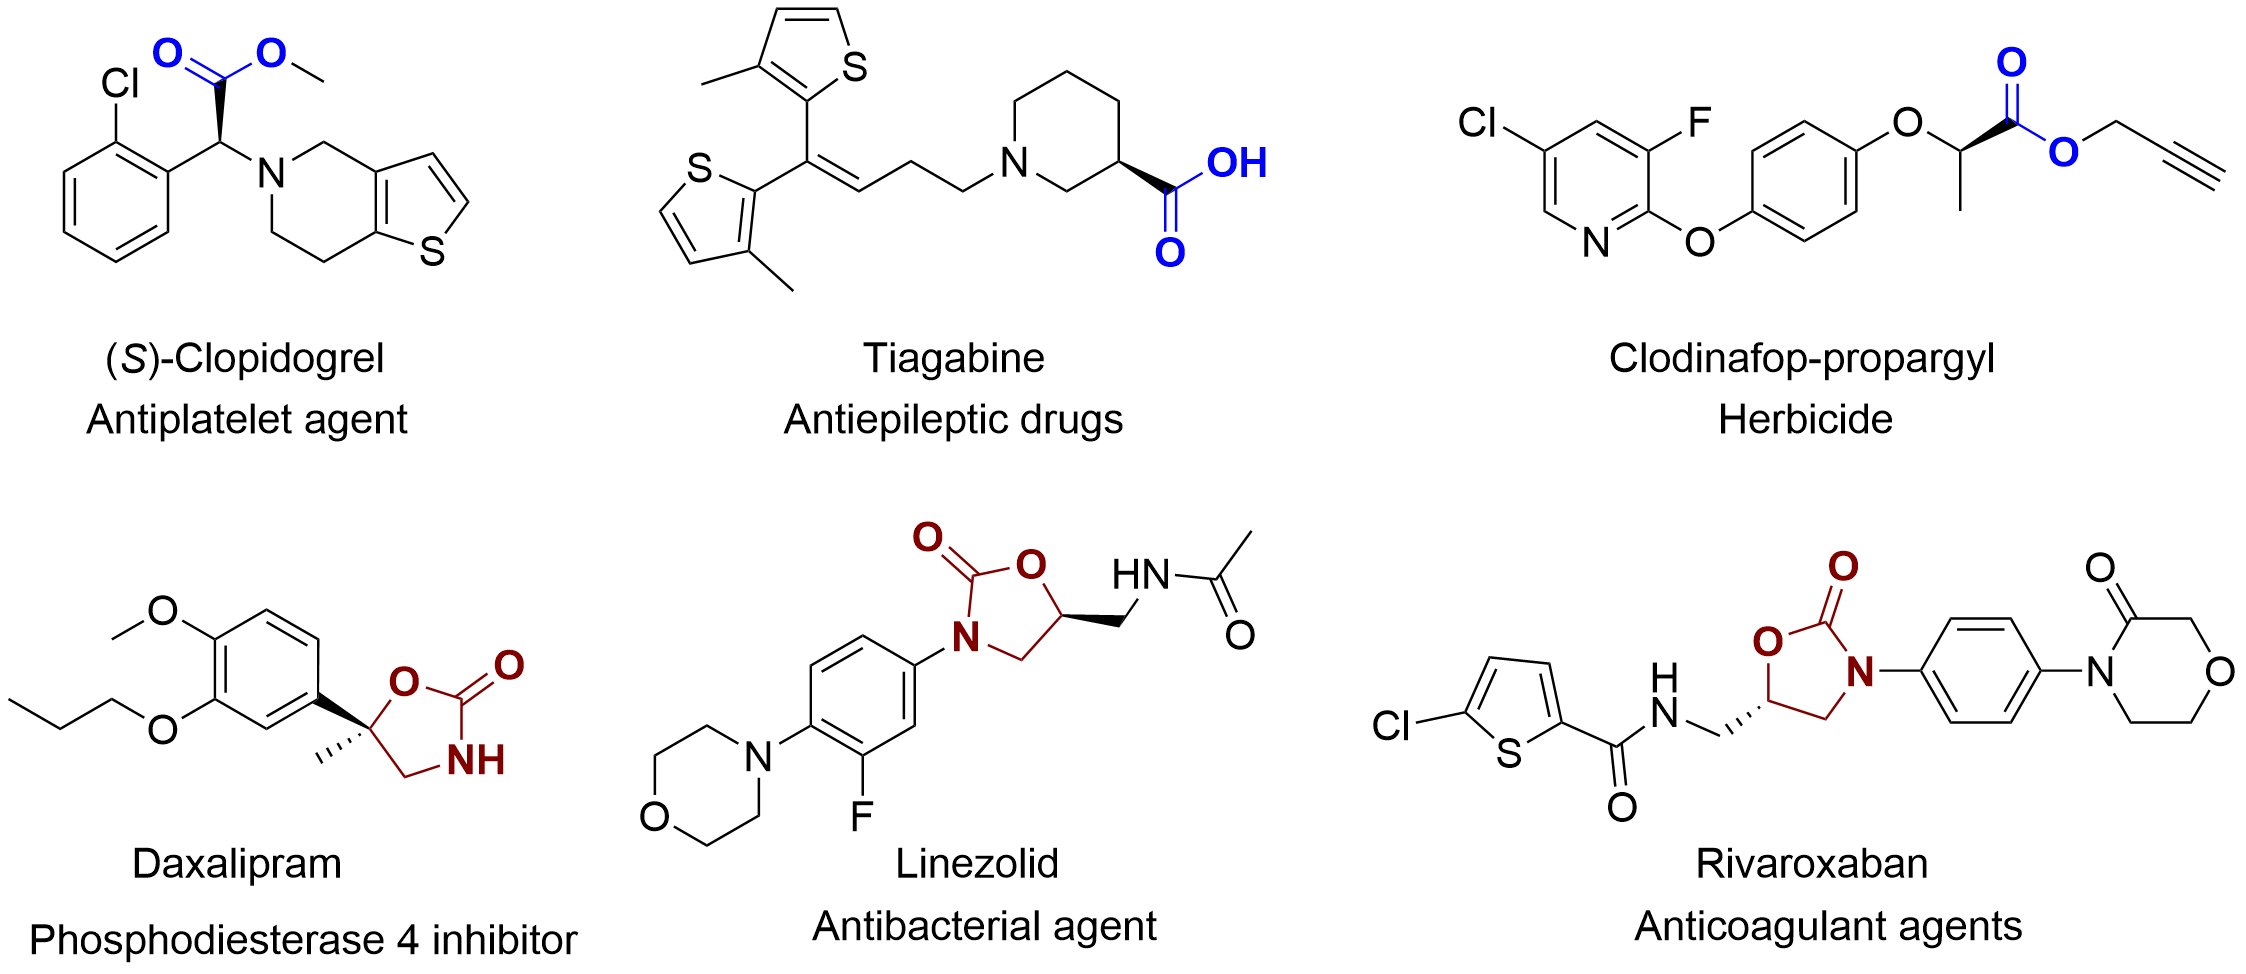
Carbon dioxide (CO2) is an abundant, inexpensive, readily available, nontoxic, and renewable C1 resource[1]. Its conversion into high-value-added chemicals provides a new material basis for sustainable chemical synthesis and represents an important pathway toward carbon resource recycling and green, low-carbon development[2]. The development of new reactions for CO2 utilization has therefore become a research focus in synthetic chemistry, among which the construction of chiral carboxylic acids and chiral heterocyclic compounds using CO2 is particularly valuable[3]. Chiral carboxylic acids constitute an important class of structural motifs widely present in natural products, pharmaceuticals, and agrochemicals[4]. Meanwhile, chiral heterocycles such as 2-oxazolidinones and cyclic carbonates serve as core scaffolds in numerous biologically active molecules and pharmaceutical agents (Scheme 1)[5]. These chiral fragments play an irreplaceable role in regulating stereoselective recognition between drug molecules and biological targets. The development of efficient and highly selective catalytic methods for the precise construction of such chiral fragments is therefore a frontier subject in synthetic chemistry and a key driver of innovation in chiral pharmaceuticals and green agrochemicals.
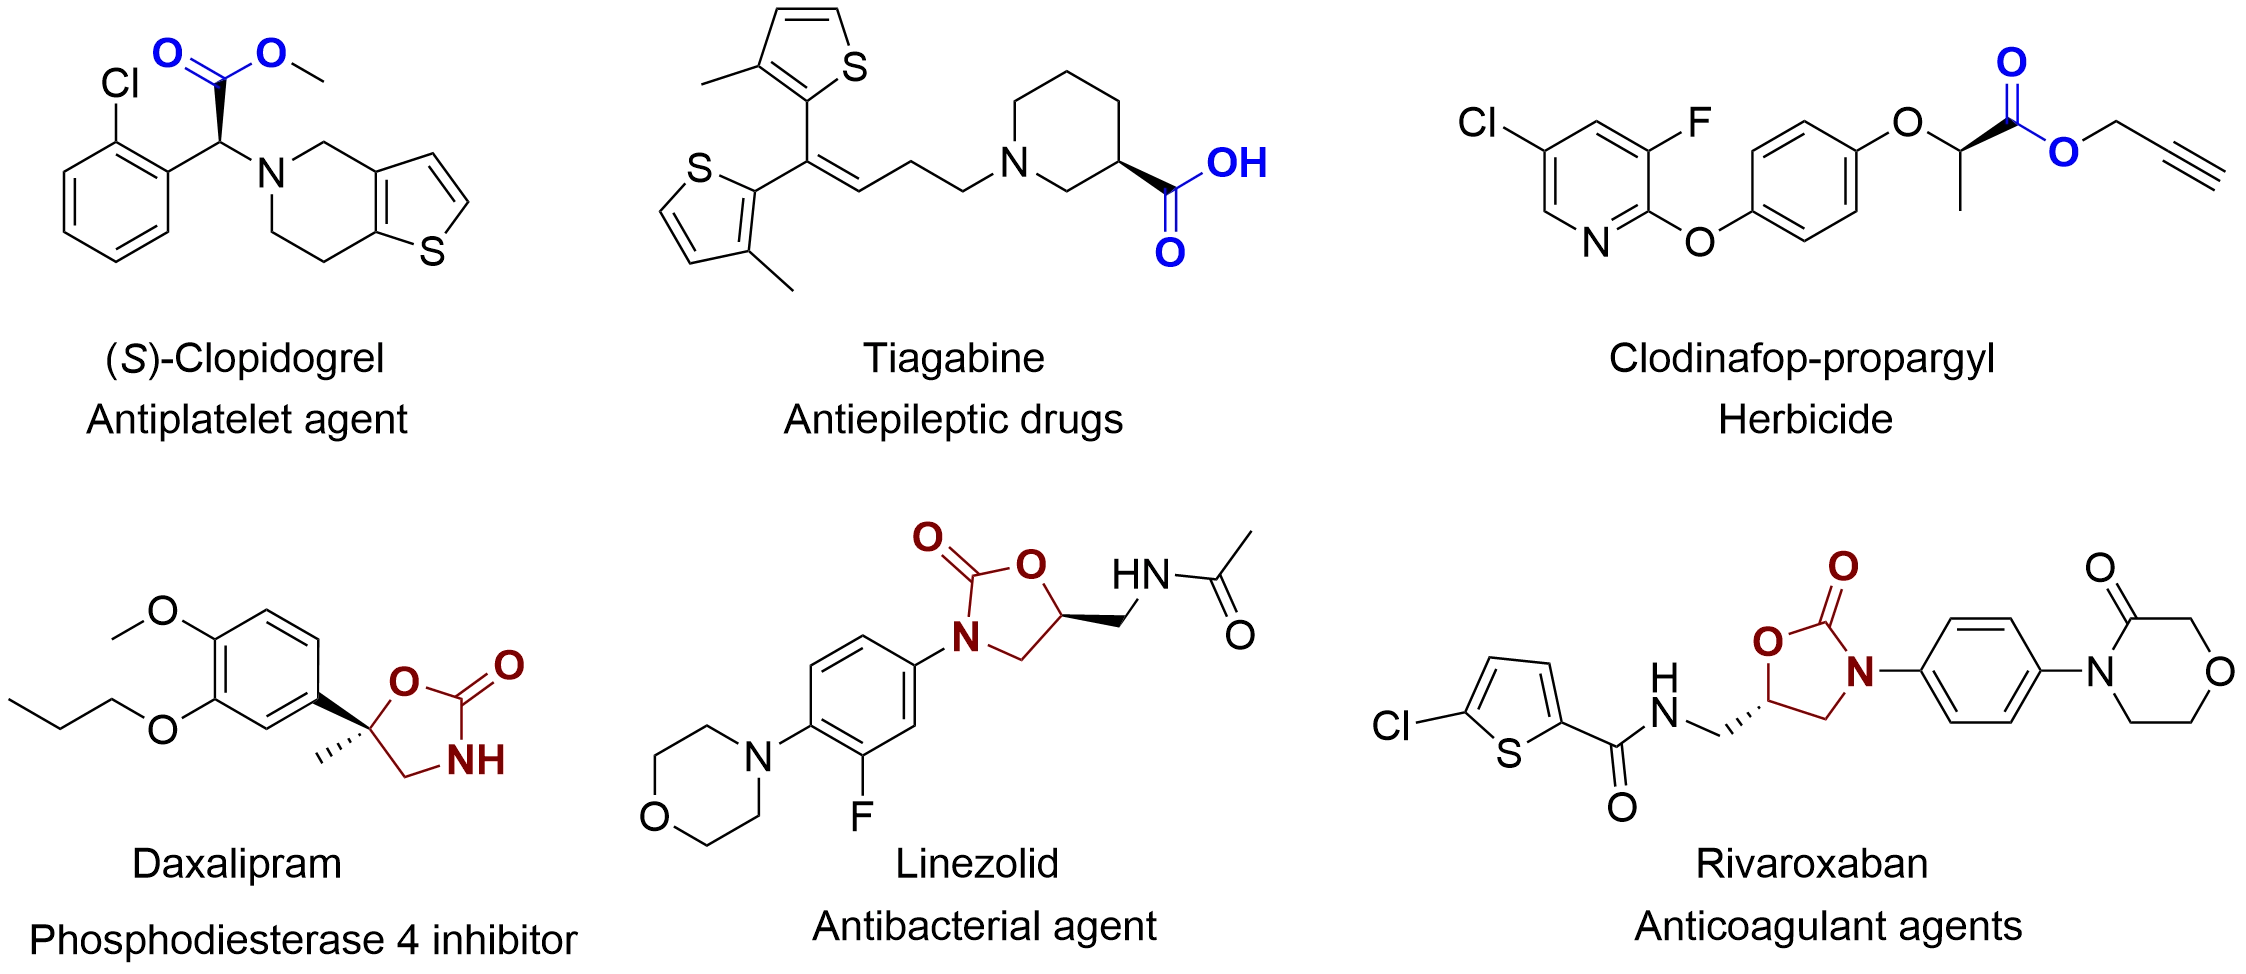
Scheme 1. Representative examples of chiral carboxylic acids and chiral heterocycles.
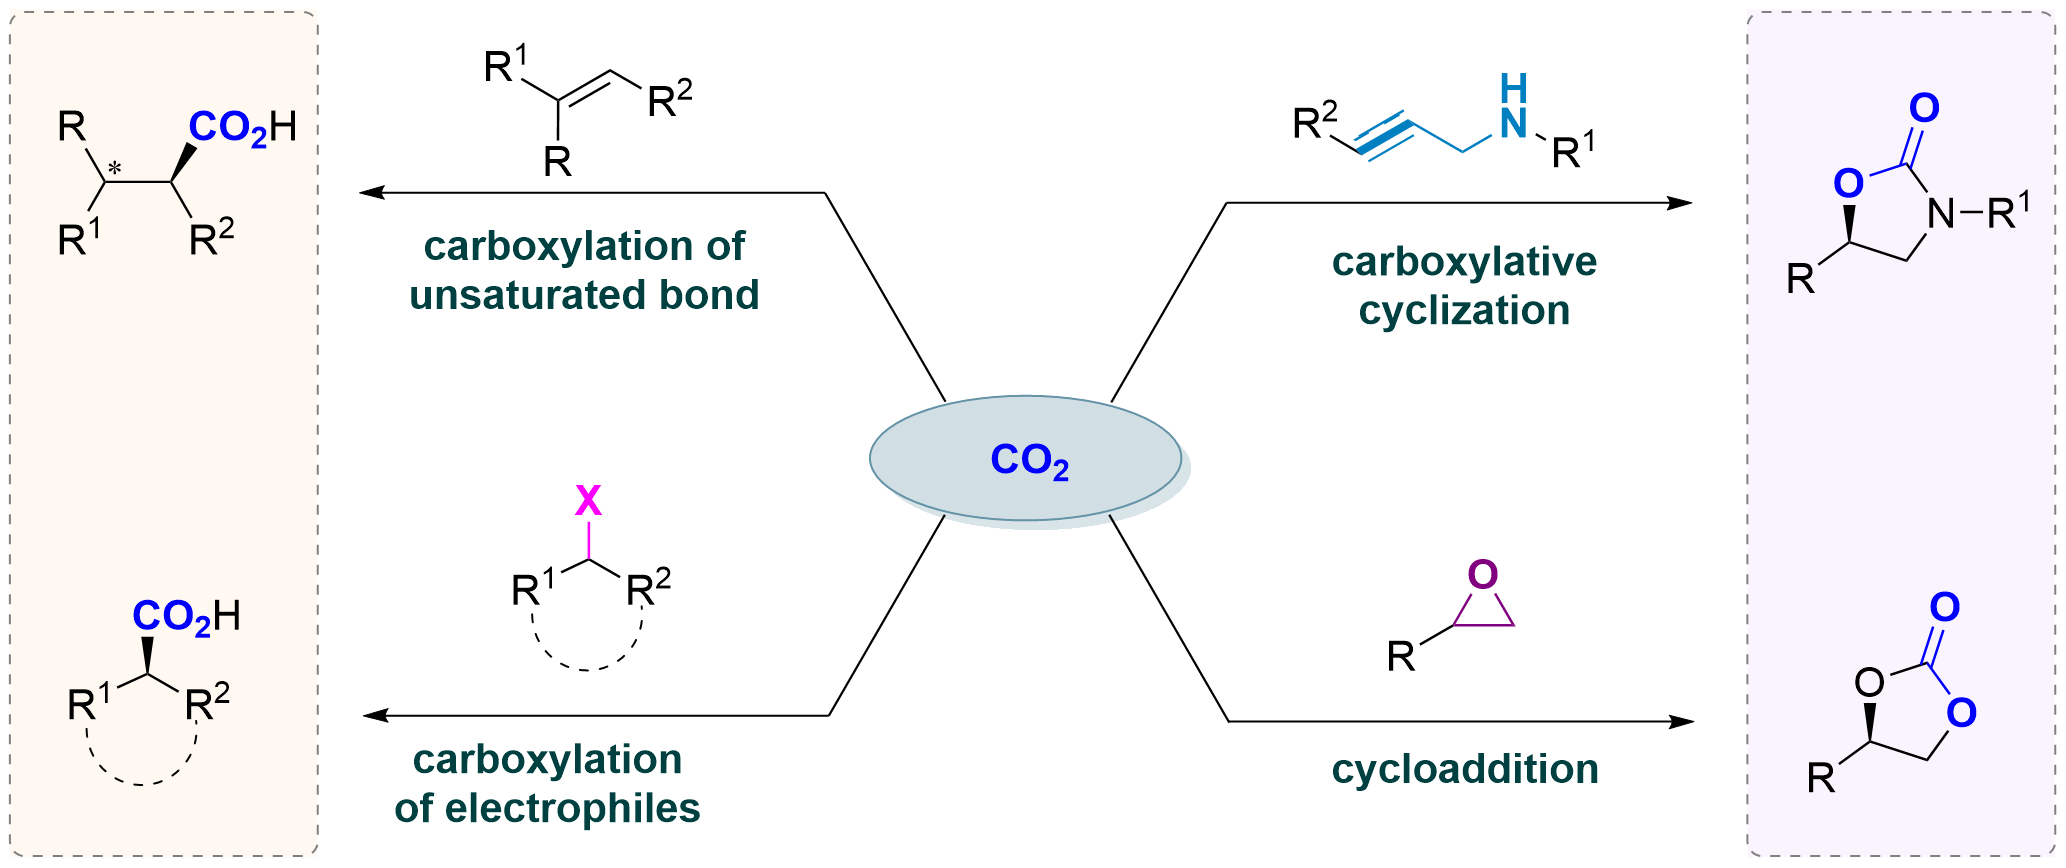
Conventionally, the synthesis of chiral carboxylic acids has relied primarily on the carbonylation of alkenes with CO[6], chiral auxiliary-induced asymmetric alkylation[7], and asymmetric catalytic hydrogenation of unsaturated carboxylic acids[8]. The synthesis of chiral heterocycles, as exemplified by chiral 2-oxazolidinones, has typically involved the cyclization of chiral amino alcohols with phosgene or its derivatives[9], the transformation of highly toxic isocyanates with glycidols and their derivatives[10], or the asymmetric hydrogenation of oxazolones[11]. For chiral cyclic carbonates, a common approach entails asymmetric dihydroxylation of alkenes followed by cyclization with a carbonyl source[12]. These methods generally suffer from cumbersome procedures, poor atom economy, limited functional group tolerance, or the requirement for prefunctionalized substrates. In contrast, employing CO2 as a C1 synthon to construct chiral carboxylic acids and chiral heterocycles through asymmetric catalysis, with simultaneous formation of carbon-carbon or carbon-heteroatom bonds, offers advantages including high atom economy, the use of green and readily available starting materials, and mild reaction conditions[13]. This strategy has attracted widespread attention and has been successfully applied to the asymmetric carboxylation of carbon-carbon unsaturated bonds and organic electrophiles, affording a variety of structurally diverse chiral carboxylic acids. In addition, CO2-involved cyclization reactions have been applied to the synthesis of heterocyclic compounds such as chiral 2-oxazolidinones and chiral cyclic carbonates (Scheme 2). However, CO2 is the end product of all carbon-based combustion processes, possesses the highest oxidation state of carbon, and is both thermodynamically stable and kinetically inert—all of which render the catalytic enantioselective fixation of CO2 a formidable challenge. With the advances in modern synthetic chemistry, significant progress has been made in catalytic enantioselective synthesis using CO2 as a C1 synthon. General reviews on enantioselective reactions with CO2 have been provided by Kleij[12b], Lu[14], Coates[15], Bayer & Hopmann[3a], Chen & Li & Xia[3d], Gui & Yu[3b], among others. However, these earlier reviews predominantly focus on literature published over five years ago, or concentrate on typical product types such as chiral cyclic carbonates and polycarbonates[16,17], carbamates[18], as well as carboxylic acids bearing central chirality[13]. Importantly, the catalytic enantioselective construction of axially chiral products has received little or no attention in these previous reviews. Given the rapid progress in this active field, a timely and comprehensive review summarizing the latest developments is therefore warranted.
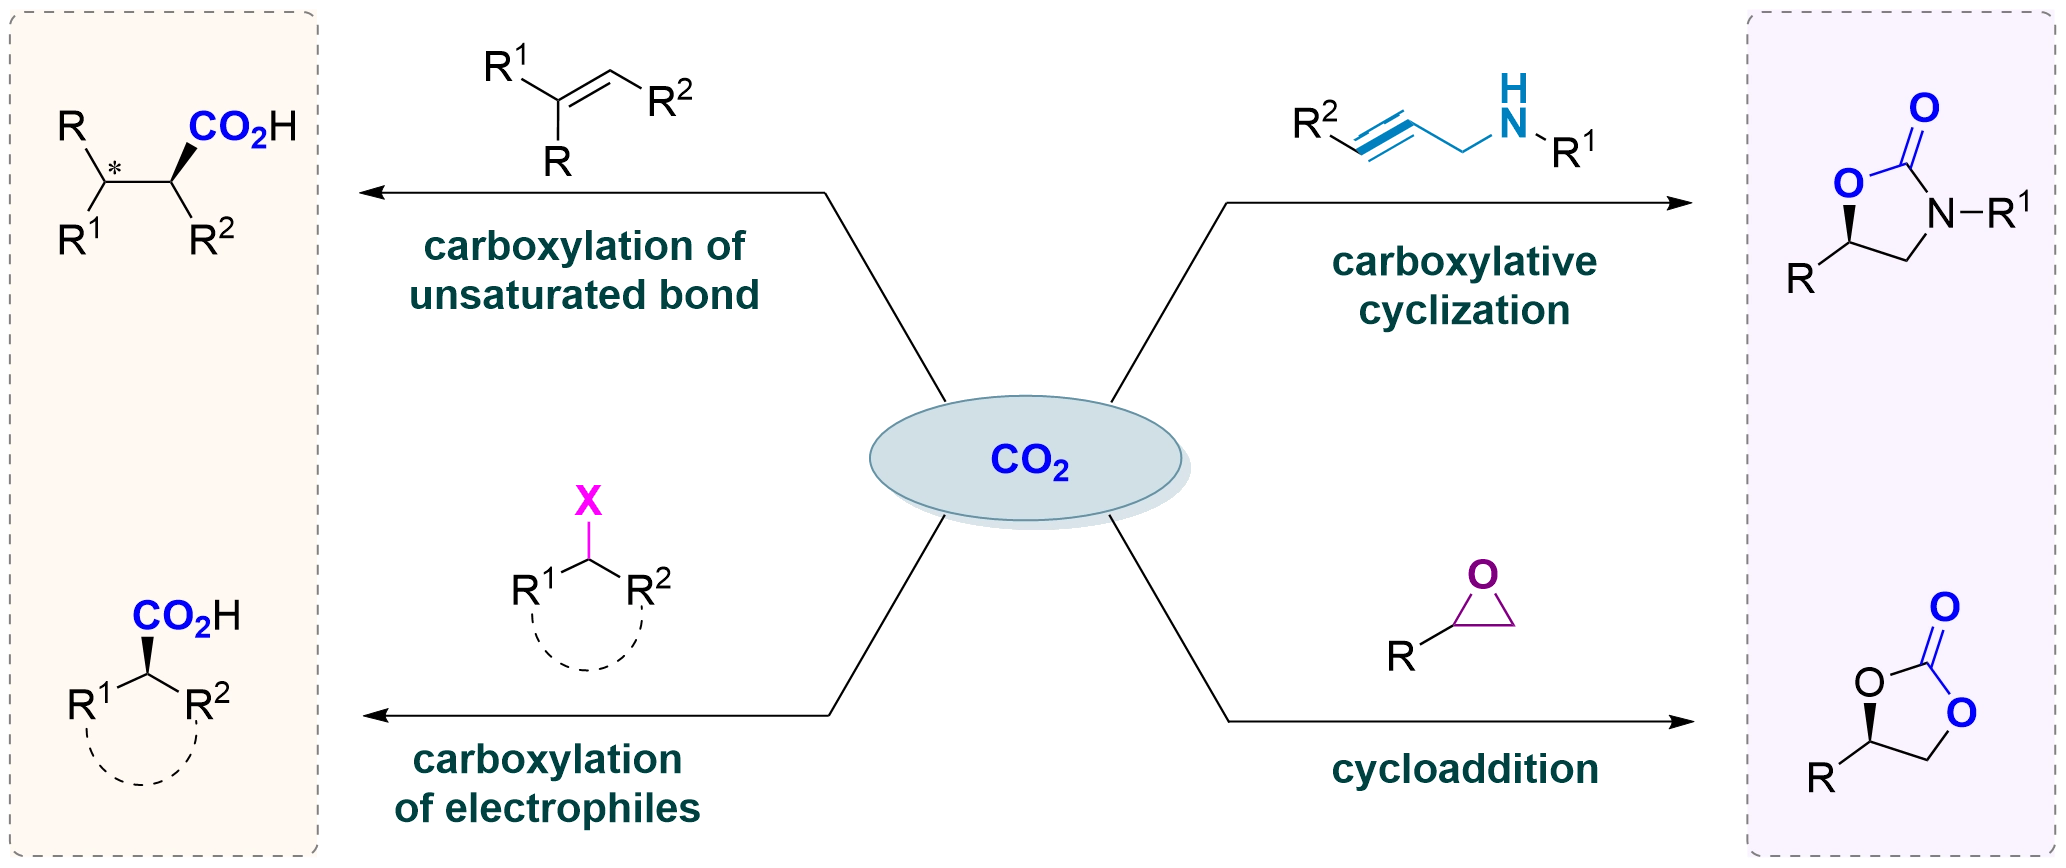
Scheme 2. Asymmetric CO₂-based synthesis of chiral carboxylic acids and heterocycles.
In this review, we systematically summarize important advances in asymmetric catalytic synthesis using CO2 as a feedstock from 2021 to mid-2026, adopting a product-oriented perspective. Emphasis is placed on strategies for achieving high enantioselectivity, with possible mechanisms and applications also discussed. The content is divided into two major sections based on target product structure: chiral carboxylic acids and chiral heterocycles. In the section on chiral carboxylic acids, the discussion is further subdivided by substrate type into the asymmetric carboxylation of carbon-carbon unsaturated bonds and organic electrophiles, with emphasis on catalytic strategies and mechanisms of stereoselectivity control. In the section on chiral heterocycles, the discussion is organized by product structure, encompassing the asymmetric synthesis of chiral 2-oxazolidinones and chiral cyclic carbonates. Finally, we summarize the major challenges and provide an outlook on future directions.
2. Asymmetric Synthesis of Chiral Carboxylic Acids Involving CO2
2.1 Asymmetric carboxylation of carbon-carbon unsaturated bonds with CO2
The asymmetric carboxylation of carbon-carbon unsaturated bonds using CO2 as the carboxyl source represents a direct and atom-economical route to chiral carboxylic acids and their derivatives. Alkenes and alkynes, as two fundamental classes of readily available unsaturated hydrocarbons, serve as ideal substrates for this strategy. However, the low reactivity of CO2 and its weak coordination ability with metals render it challenging to precisely control the regio- and stereoselectivity of CO2 addition to unsaturated bonds within the catalytic cycle. In recent years, through the judicious design of transition metal catalysts and chiral ligands, chemists have achieved significant breakthroughs in this field. For alkenes, four asymmetric catalytic systems, namely, carbo-carboxylation, boracarboxylation, hydrocarboxylation, and dicarboxylation, have been established; for alkynes, asymmetric hydroxymethylation and carbo-carboxylation employing CO2 as a C1 source has been developed as the successful transformation. These systems are discussed separately below.
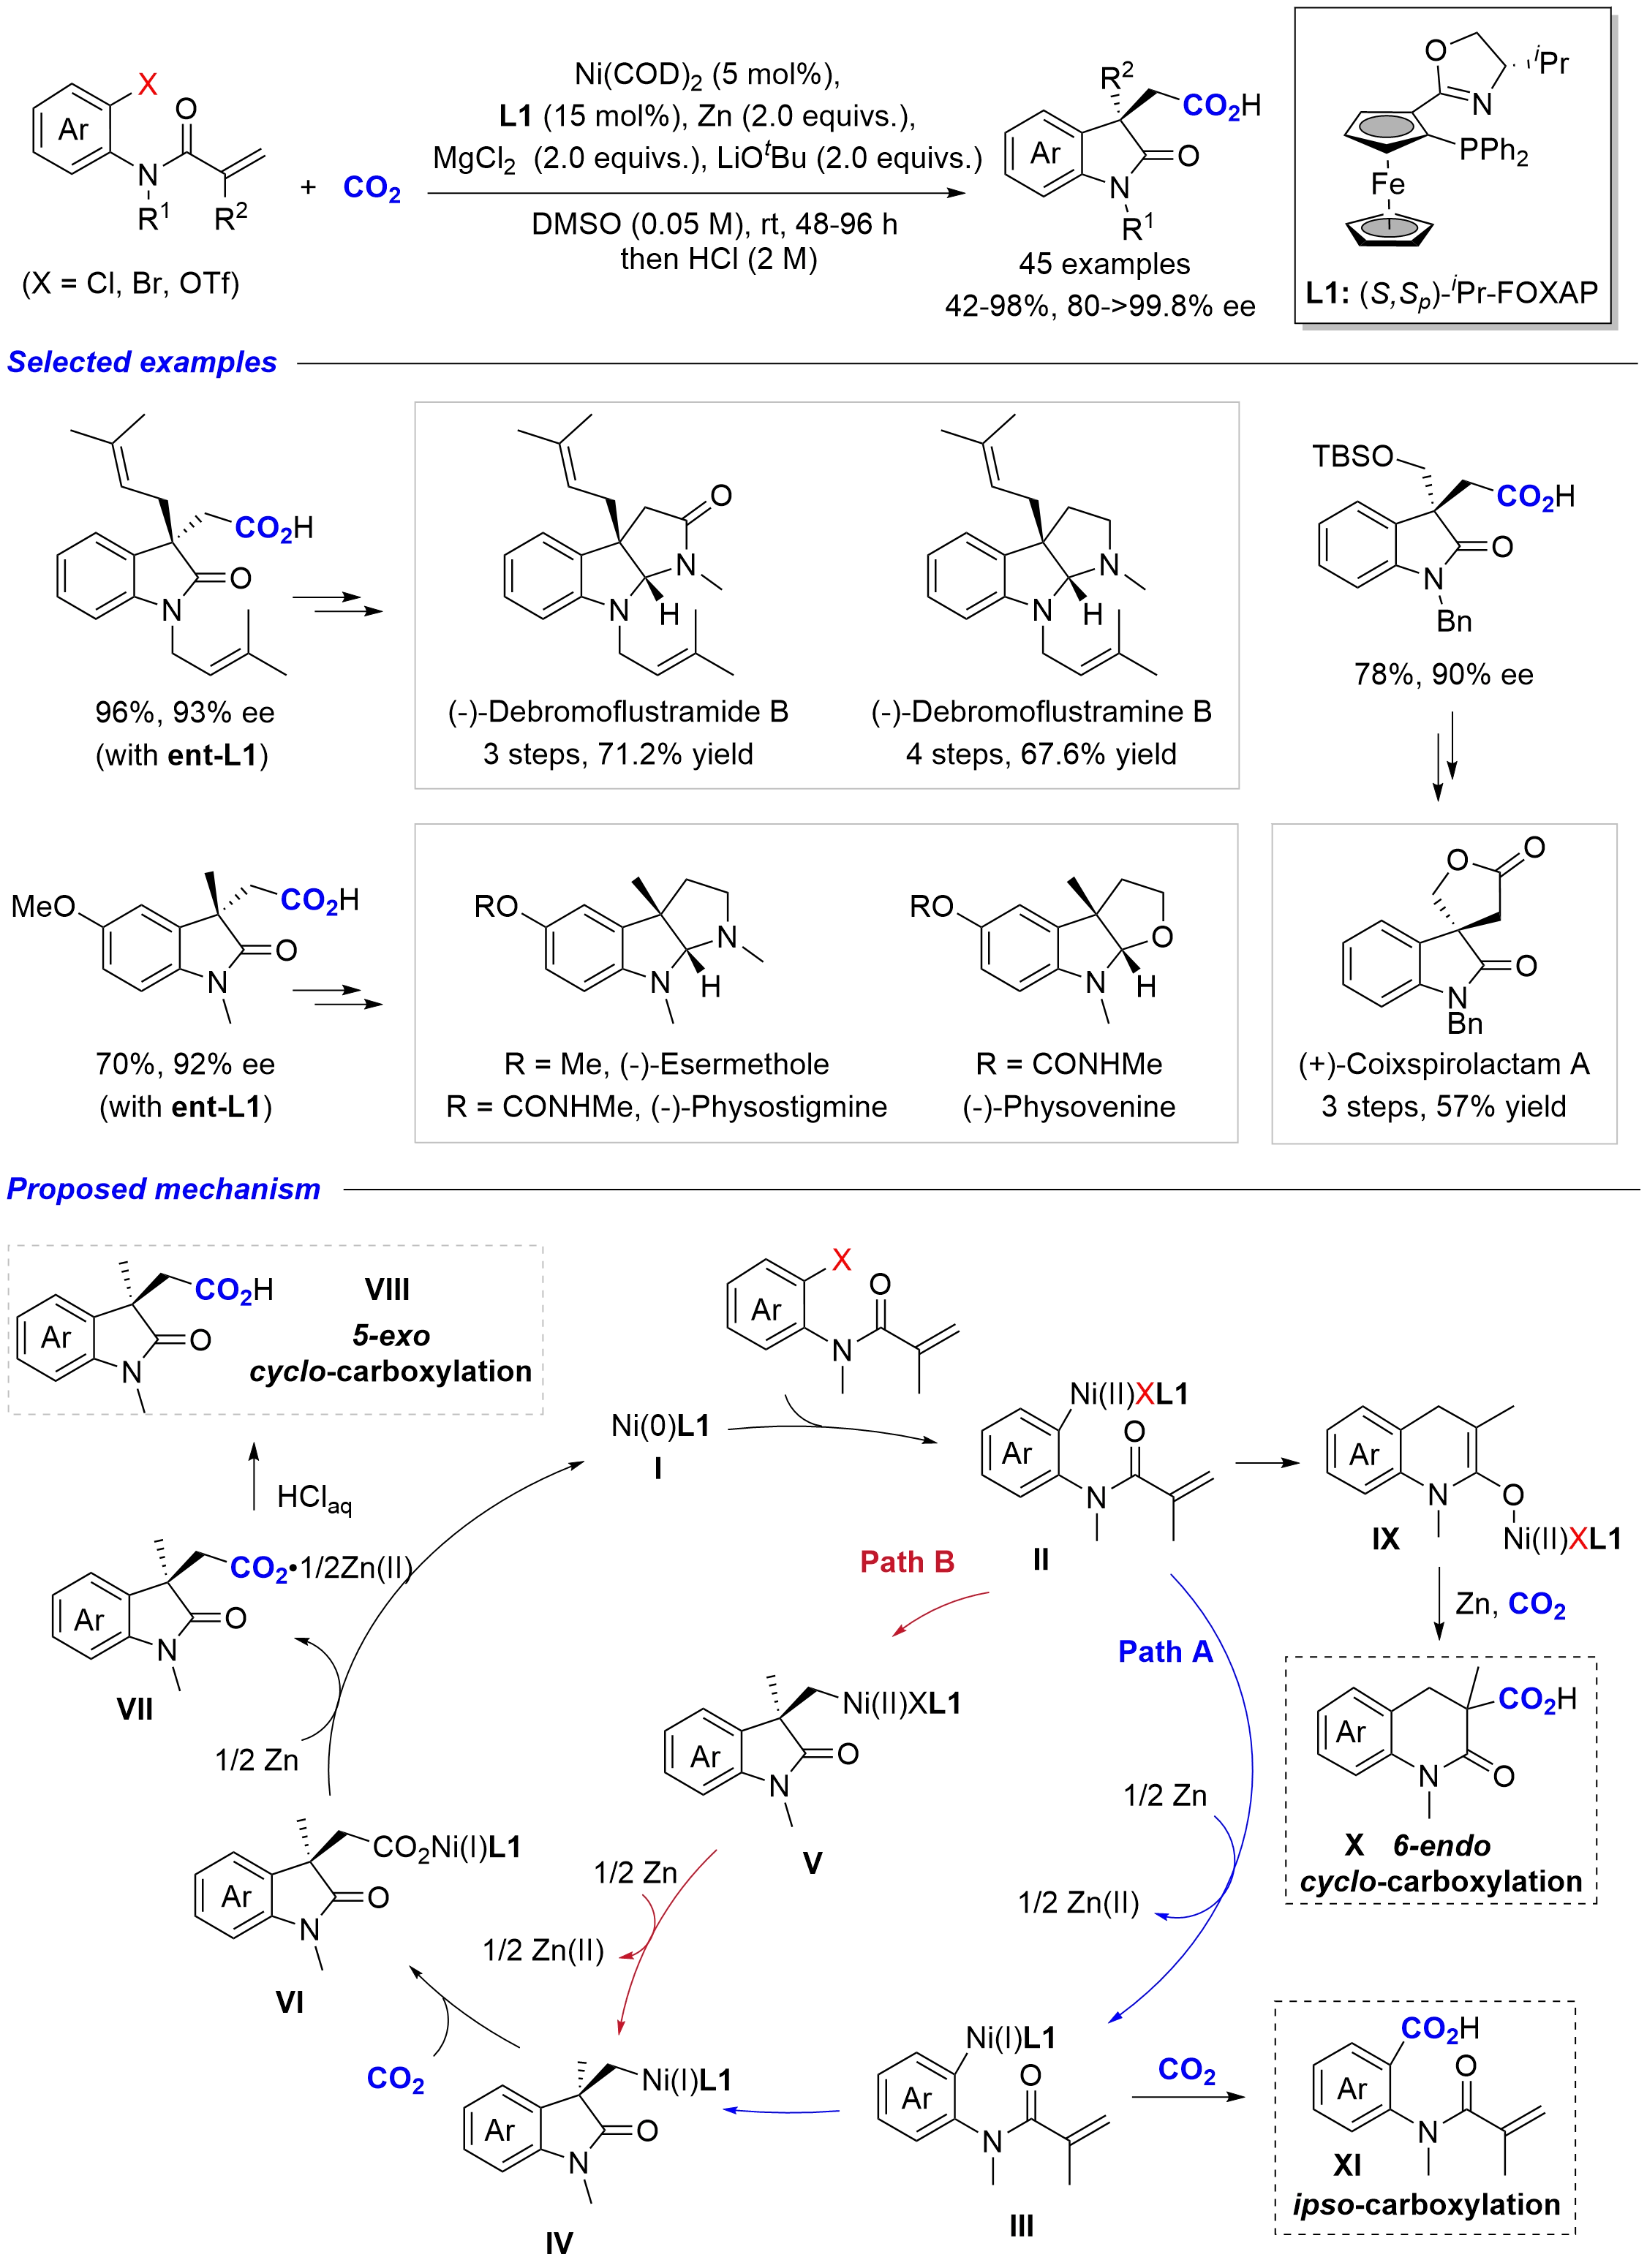
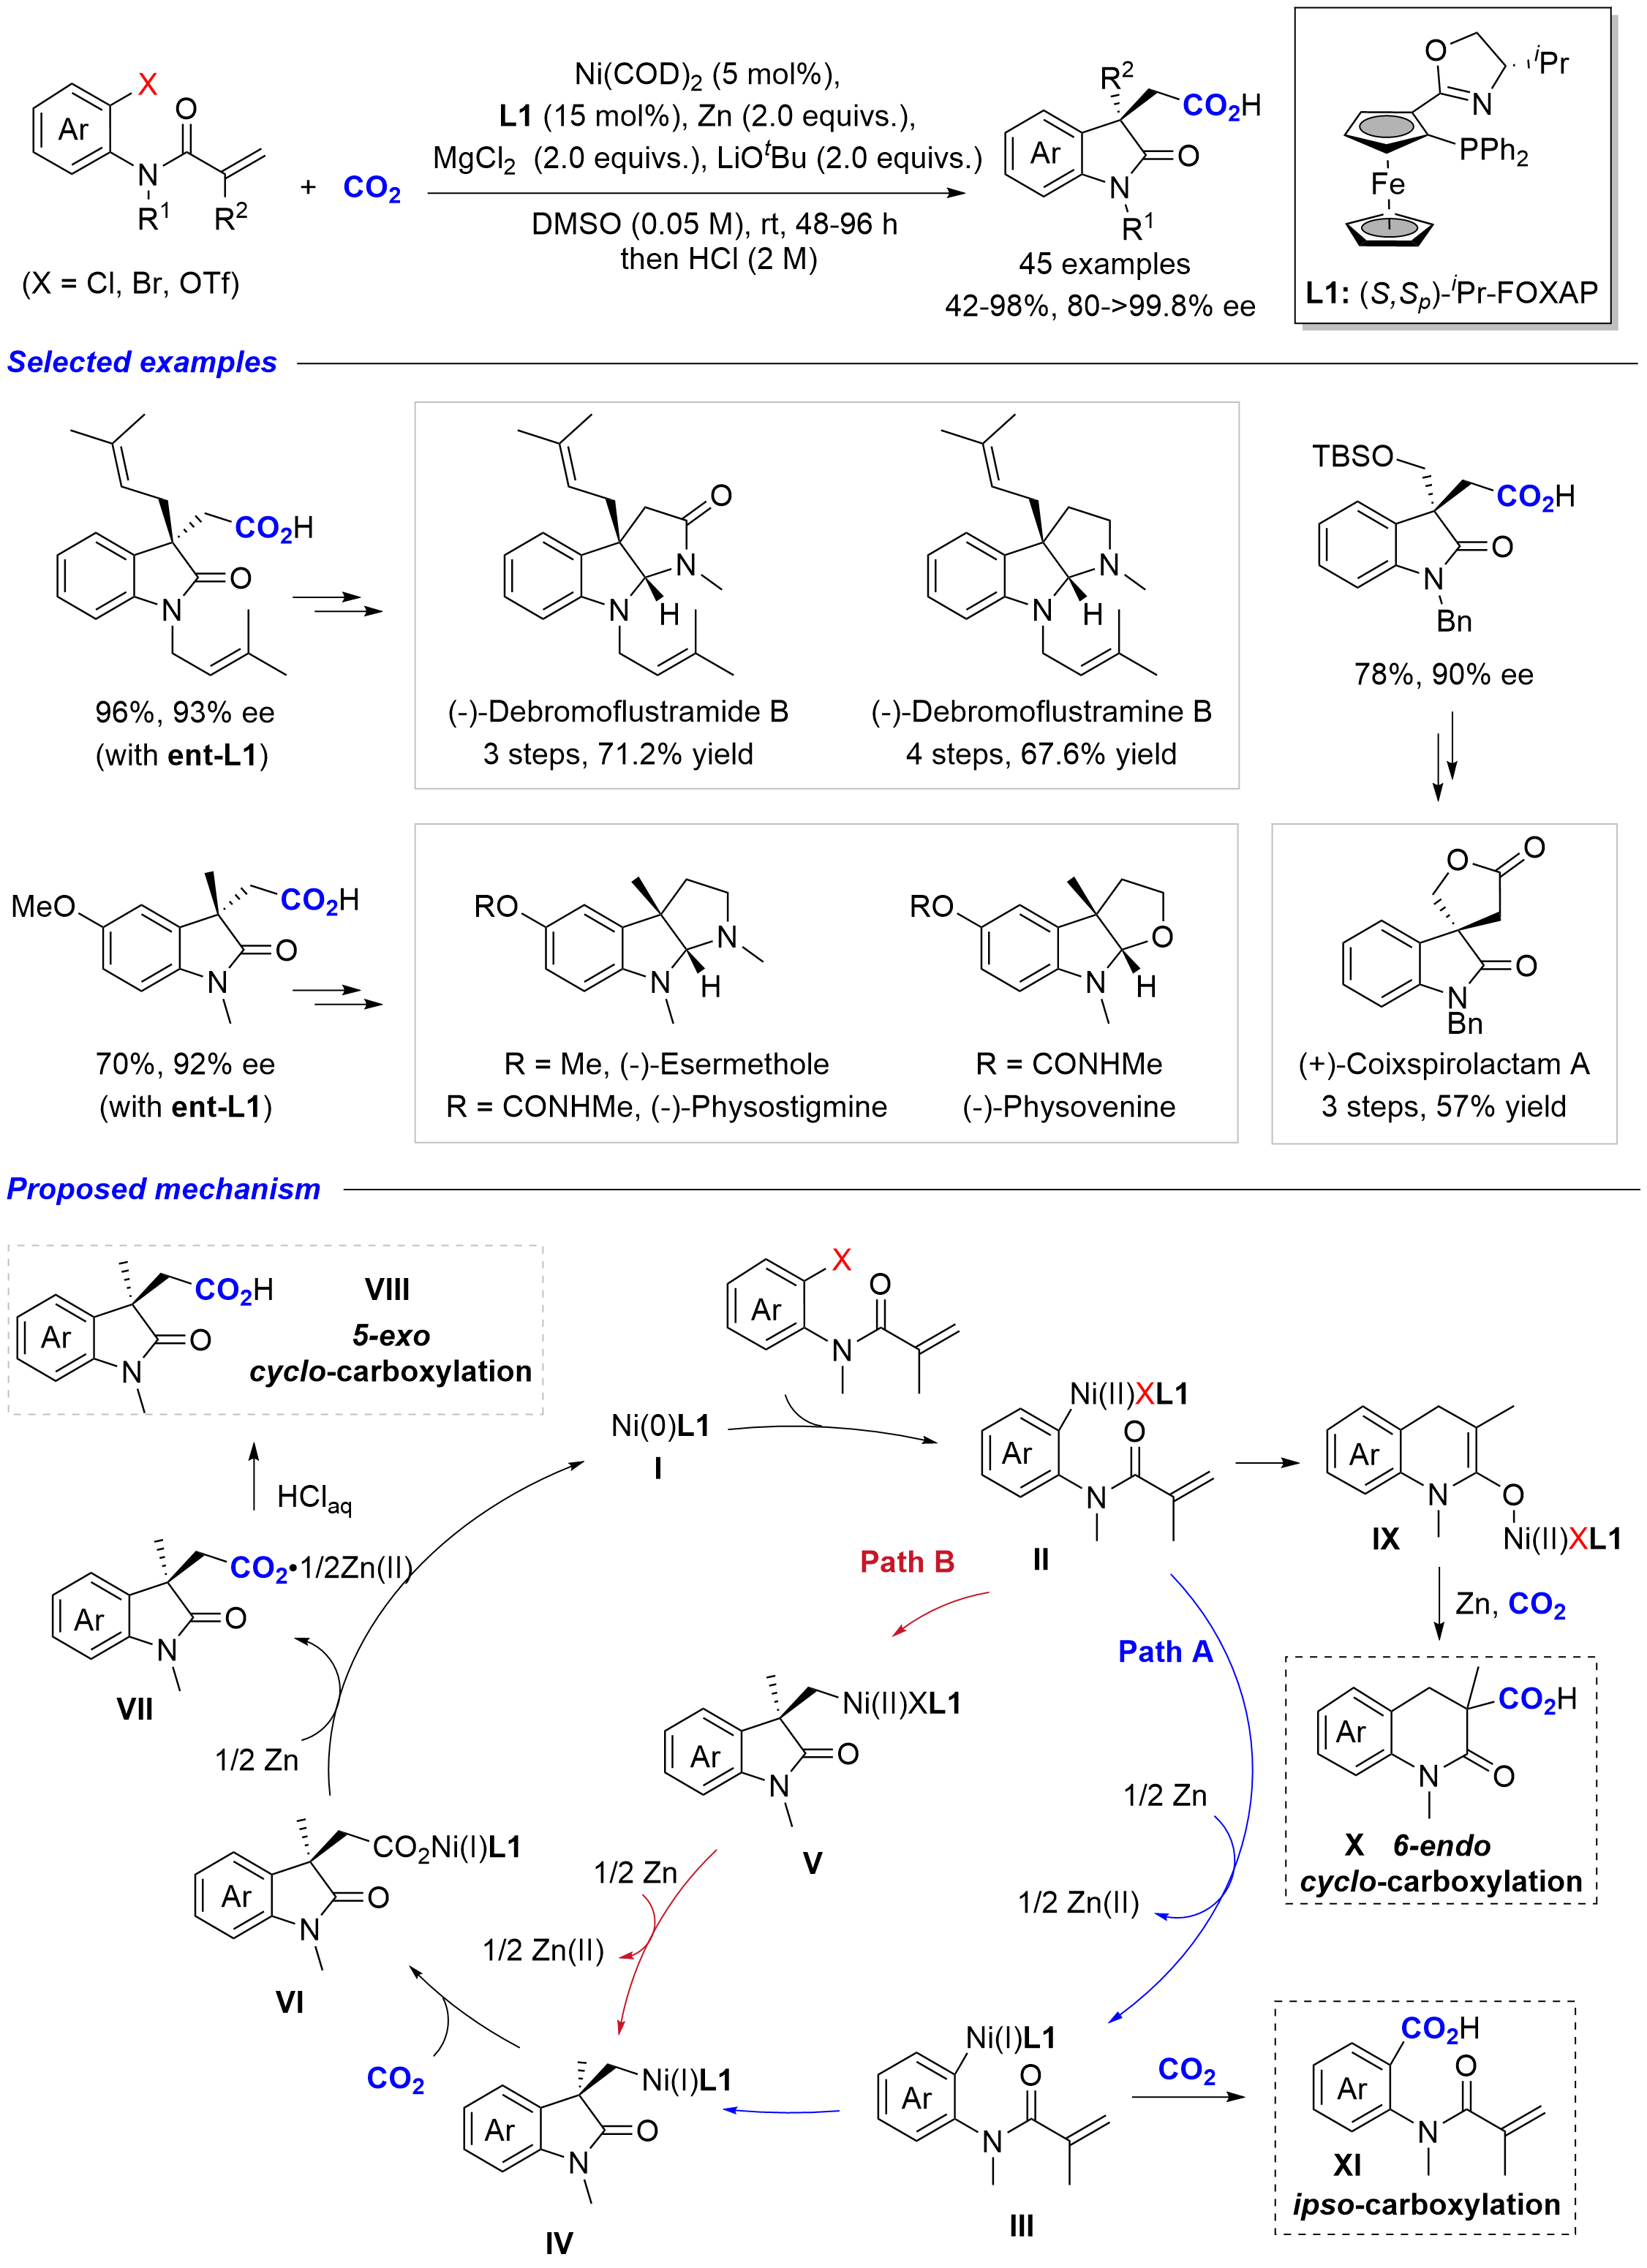
Carbo-carboxylation of alkenes. In 2021, Yu, Kong and coworkers reported the first example of a chiral nickel-catalyzed asymmetric reductive carbocarboxylation of alkenes with CO2 (Scheme 3)[19]. A variety of aryl (pseudo)halides, such as aryl bromides, aryl triflates, and inert aryl chlorides, all worked well under mild conditions, affording oxindole-3-acetic acid derivatives bearing chiral quaternary carbon stereocenters with high chemo-, regio-, and enantioselectivity. This method was successfully applied to the total synthesis of (-)-Debromoflustramide B, (-)-Debromoflustramine B, and (+)-Coixspirolactam A, as well as the formal synthesis of (-)-Esermethole, (-)-Physostigmine, and (-)-Physovenine. Mechanistic studies revealed that the Ni(0) species I first undergoes oxidative addition with an aryl halide to generate the Ar-Ni(II) intermediate II. This intermediate can be converted into the target product via two pathways. In Path A, II undergoes single-electron reduction by zinc powder to afford the Ar-Ni(I) species III, which then undergoes enantioselective 5-exo cyclization to furnish the alkyl-Ni(II) intermediate IV. In Path B, II can also undergo direct 5-exo cyclization to generate the alkyl-Ni(II) intermediate V, which is subsequently reduced by zinc to give IV. Intermediate IV then undergoes nucleophilic attack on CO2 to afford the Ni(I) carboxylate VI. Further reduction and transmetalation yield the carboxylate salt VII and regenerate the Ni(0) species I. Final acidic workup affords the target product VIII. In addition, the Ar-Ni(II) intermediate II may also undergo 6-endo cyclization to generate the relatively stable Ni(II) enolate IX, leading to the formation of byproduct X.
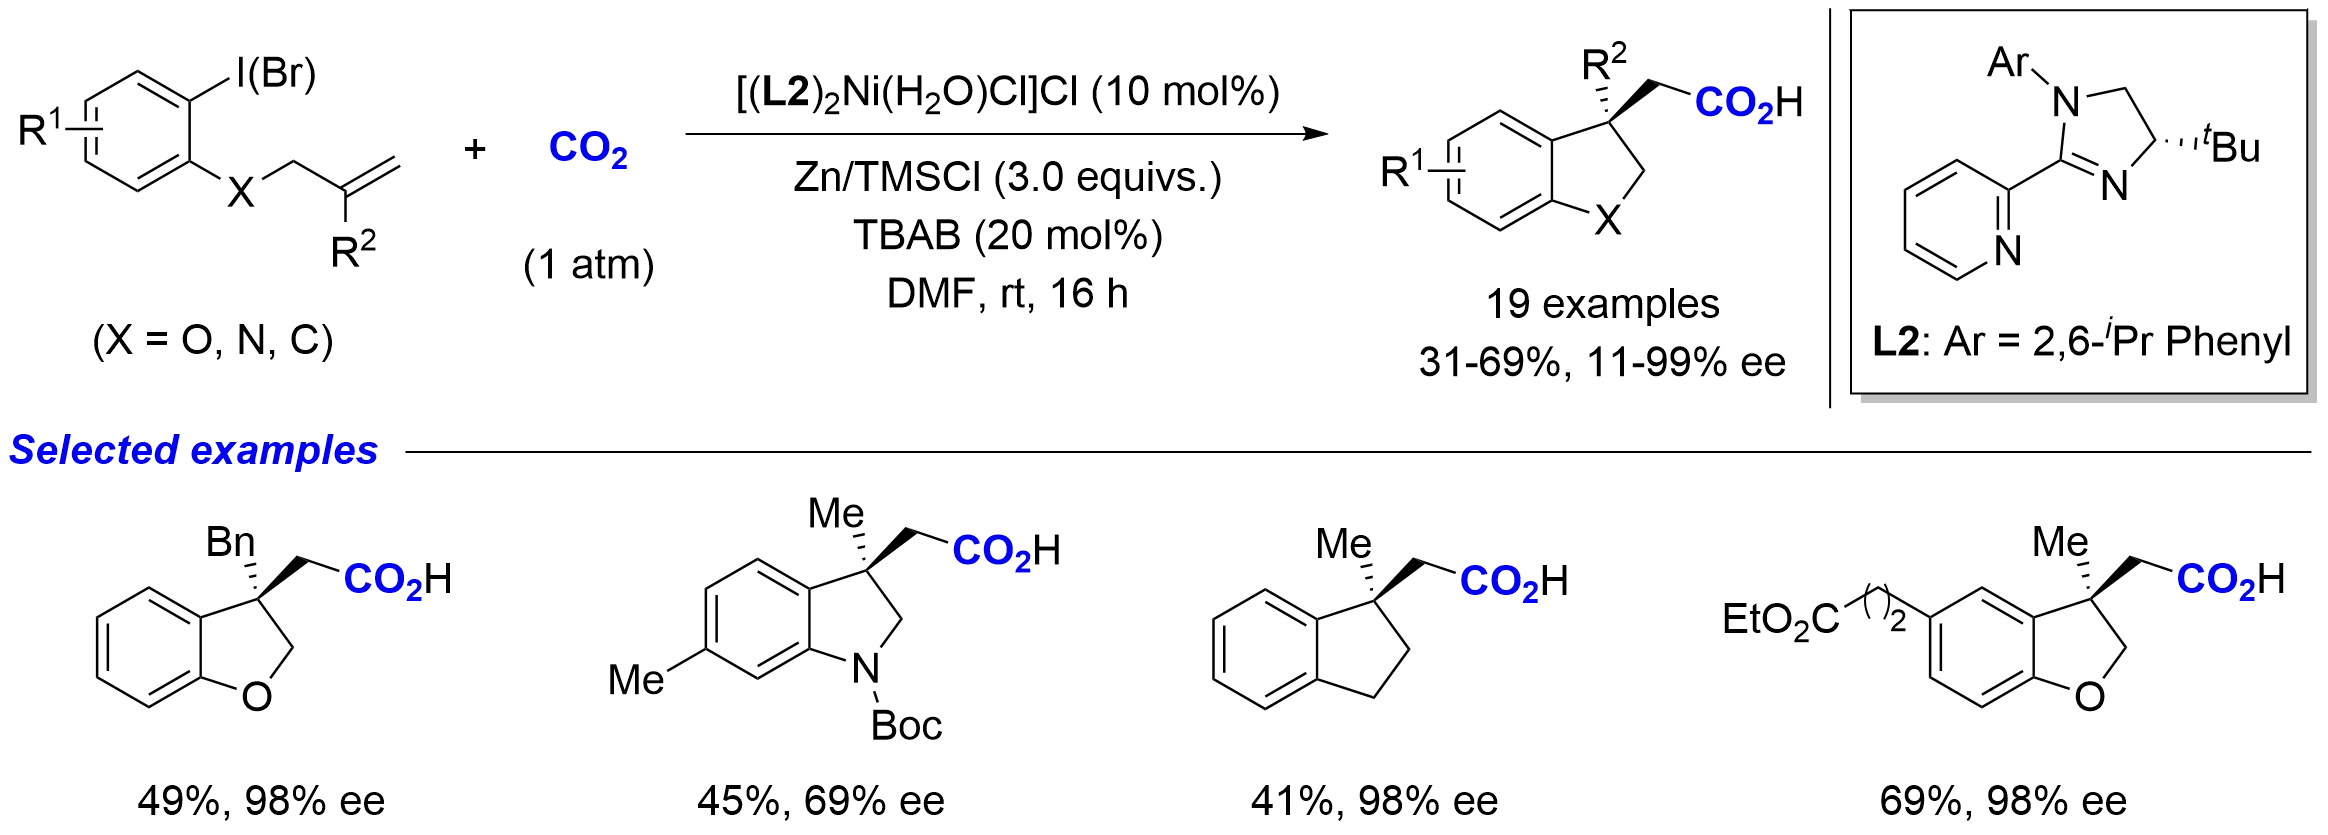
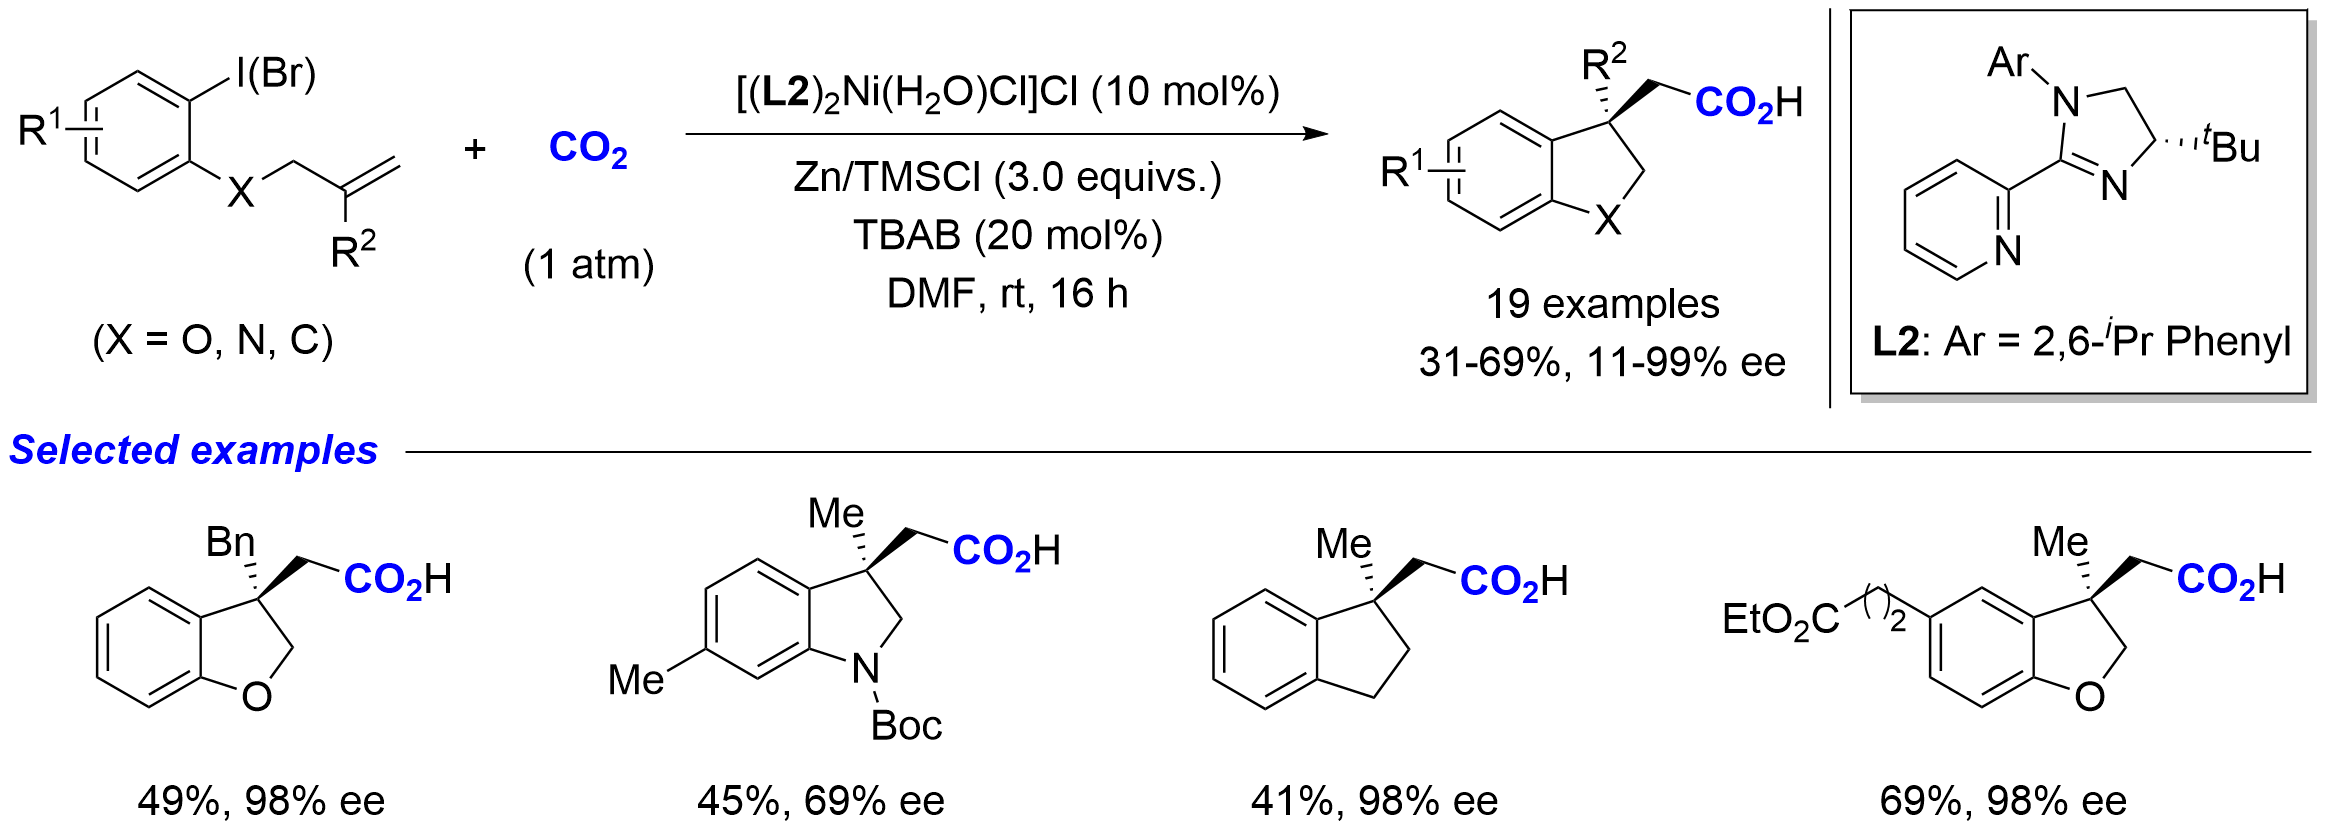
In the same year, Bandini, Silva López and coworkers reported a nickel-catalyzed enantioselective tandem Heck-carboxylation reaction, employing ortho-haloaryl allyl ethers as substrates to construct chiral 2,3-dihydrobenzofuran-3-ylacetic acids (Scheme 4)[20]. Through the design of a novel chiral pyridyl imidazoline ligand L2, the desired products were obtained with enantioselectivities up to 99% ee, and a variety of substituents on the aromatic ring as well as at the quaternary carbon center were tolerated. Distinct from the aforementioned cyclization mode, this intramolecular Heck-carboxylation cascade expanded the product scaffold diversity of nickel-catalyzed asymmetric carboxylation with CO2. DFT calculations further supported the involvement of Ni(I) species in the reaction pathway, providing mechanistic insights into the oxidation-state regulation in reductive coupling-carboxylation processes.
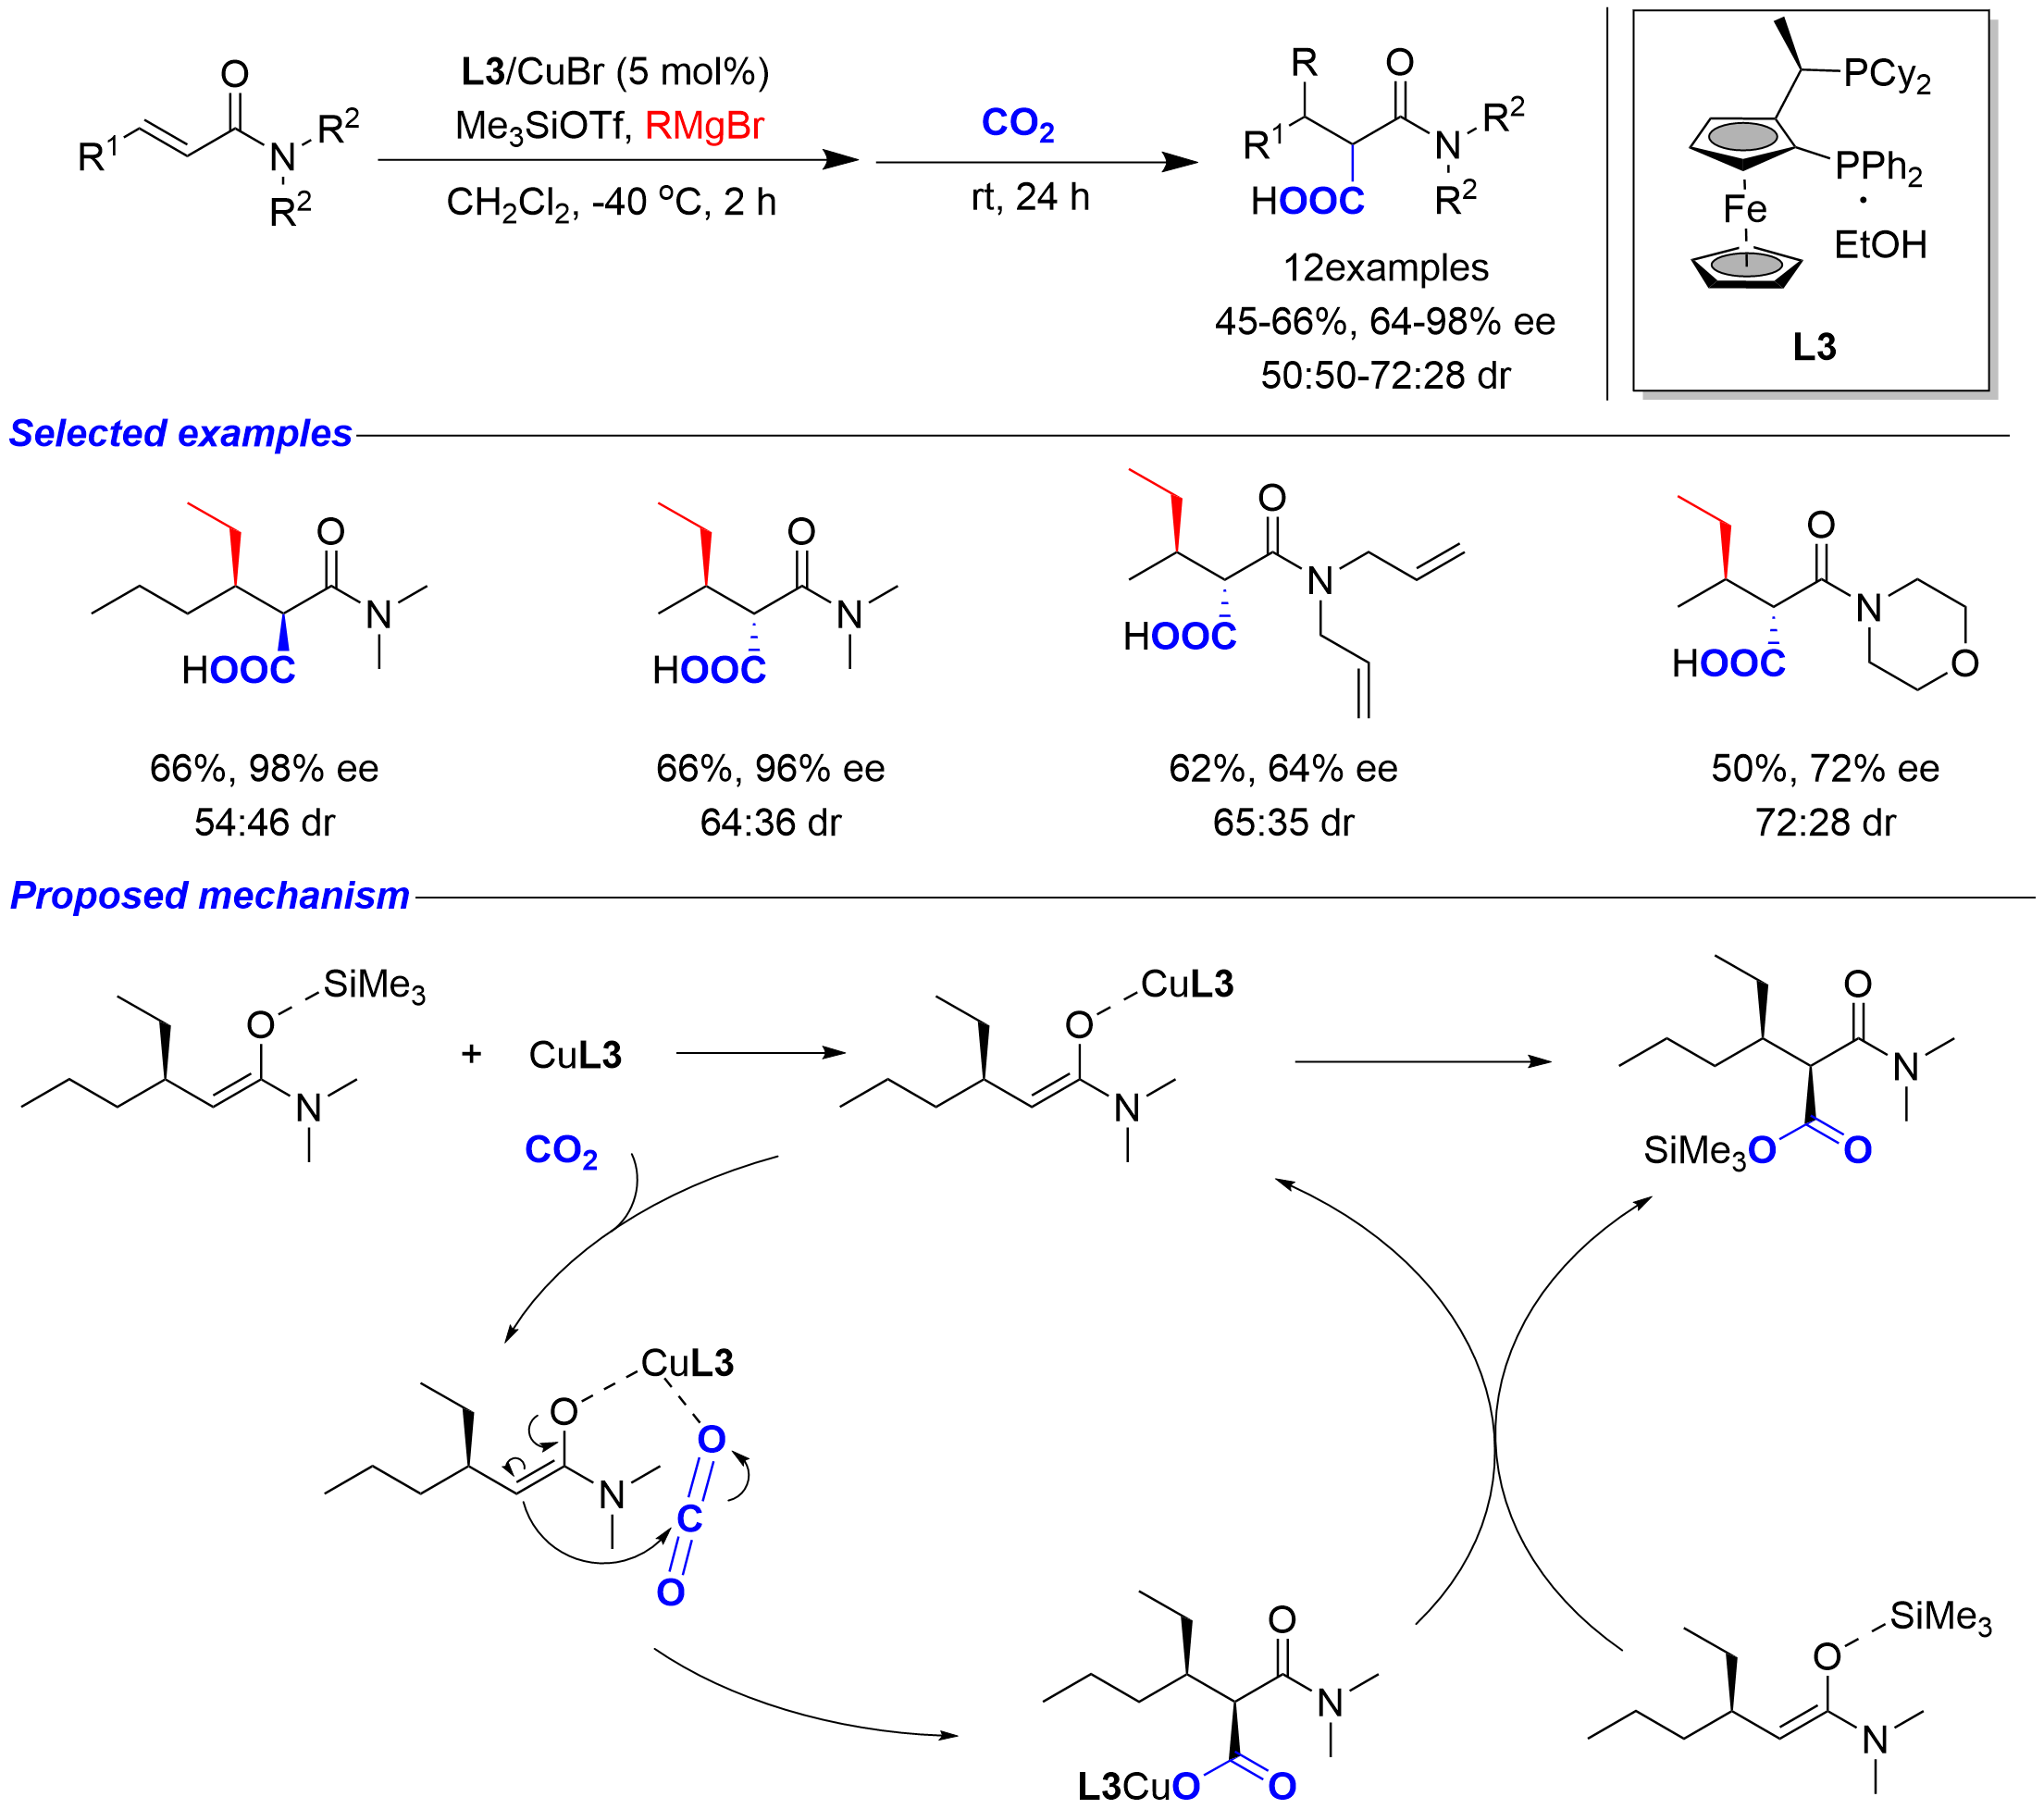
In 2024, Yan, Pan, Liu, Hong and coworkers reported a copper-catalyzed tandem asymmetric conjugate addition/CO2 capture reaction of Grignard reagents with α,β-unsaturated amides, affording chiral α-carboxyl amides bearing contiguous stereocenters in up to 66% yield, 72:28 dr and 98% ee (Scheme 5)[21]. Subsequent reduction of these α-carboxyl amides furnished the corresponding γ-amino alcohols. The reaction proceeds via enantioselective addition of a Grignard reagent to generate a chiral silyl enol ether, which is subsequently trapped by CO2 under atmospheric pressure to give the α-carboxyl amides. A range of N-substituted amides and Grignard reagents, including linear, branched, phenyl-containing, and alkenyl variants, were tolerated. It was proposed that the chiral copper catalyst exhibits a bifunctional feature, not only catalyzing the asymmetric conjugate addition but also enabling the coupling between silyl enolates and CO2.
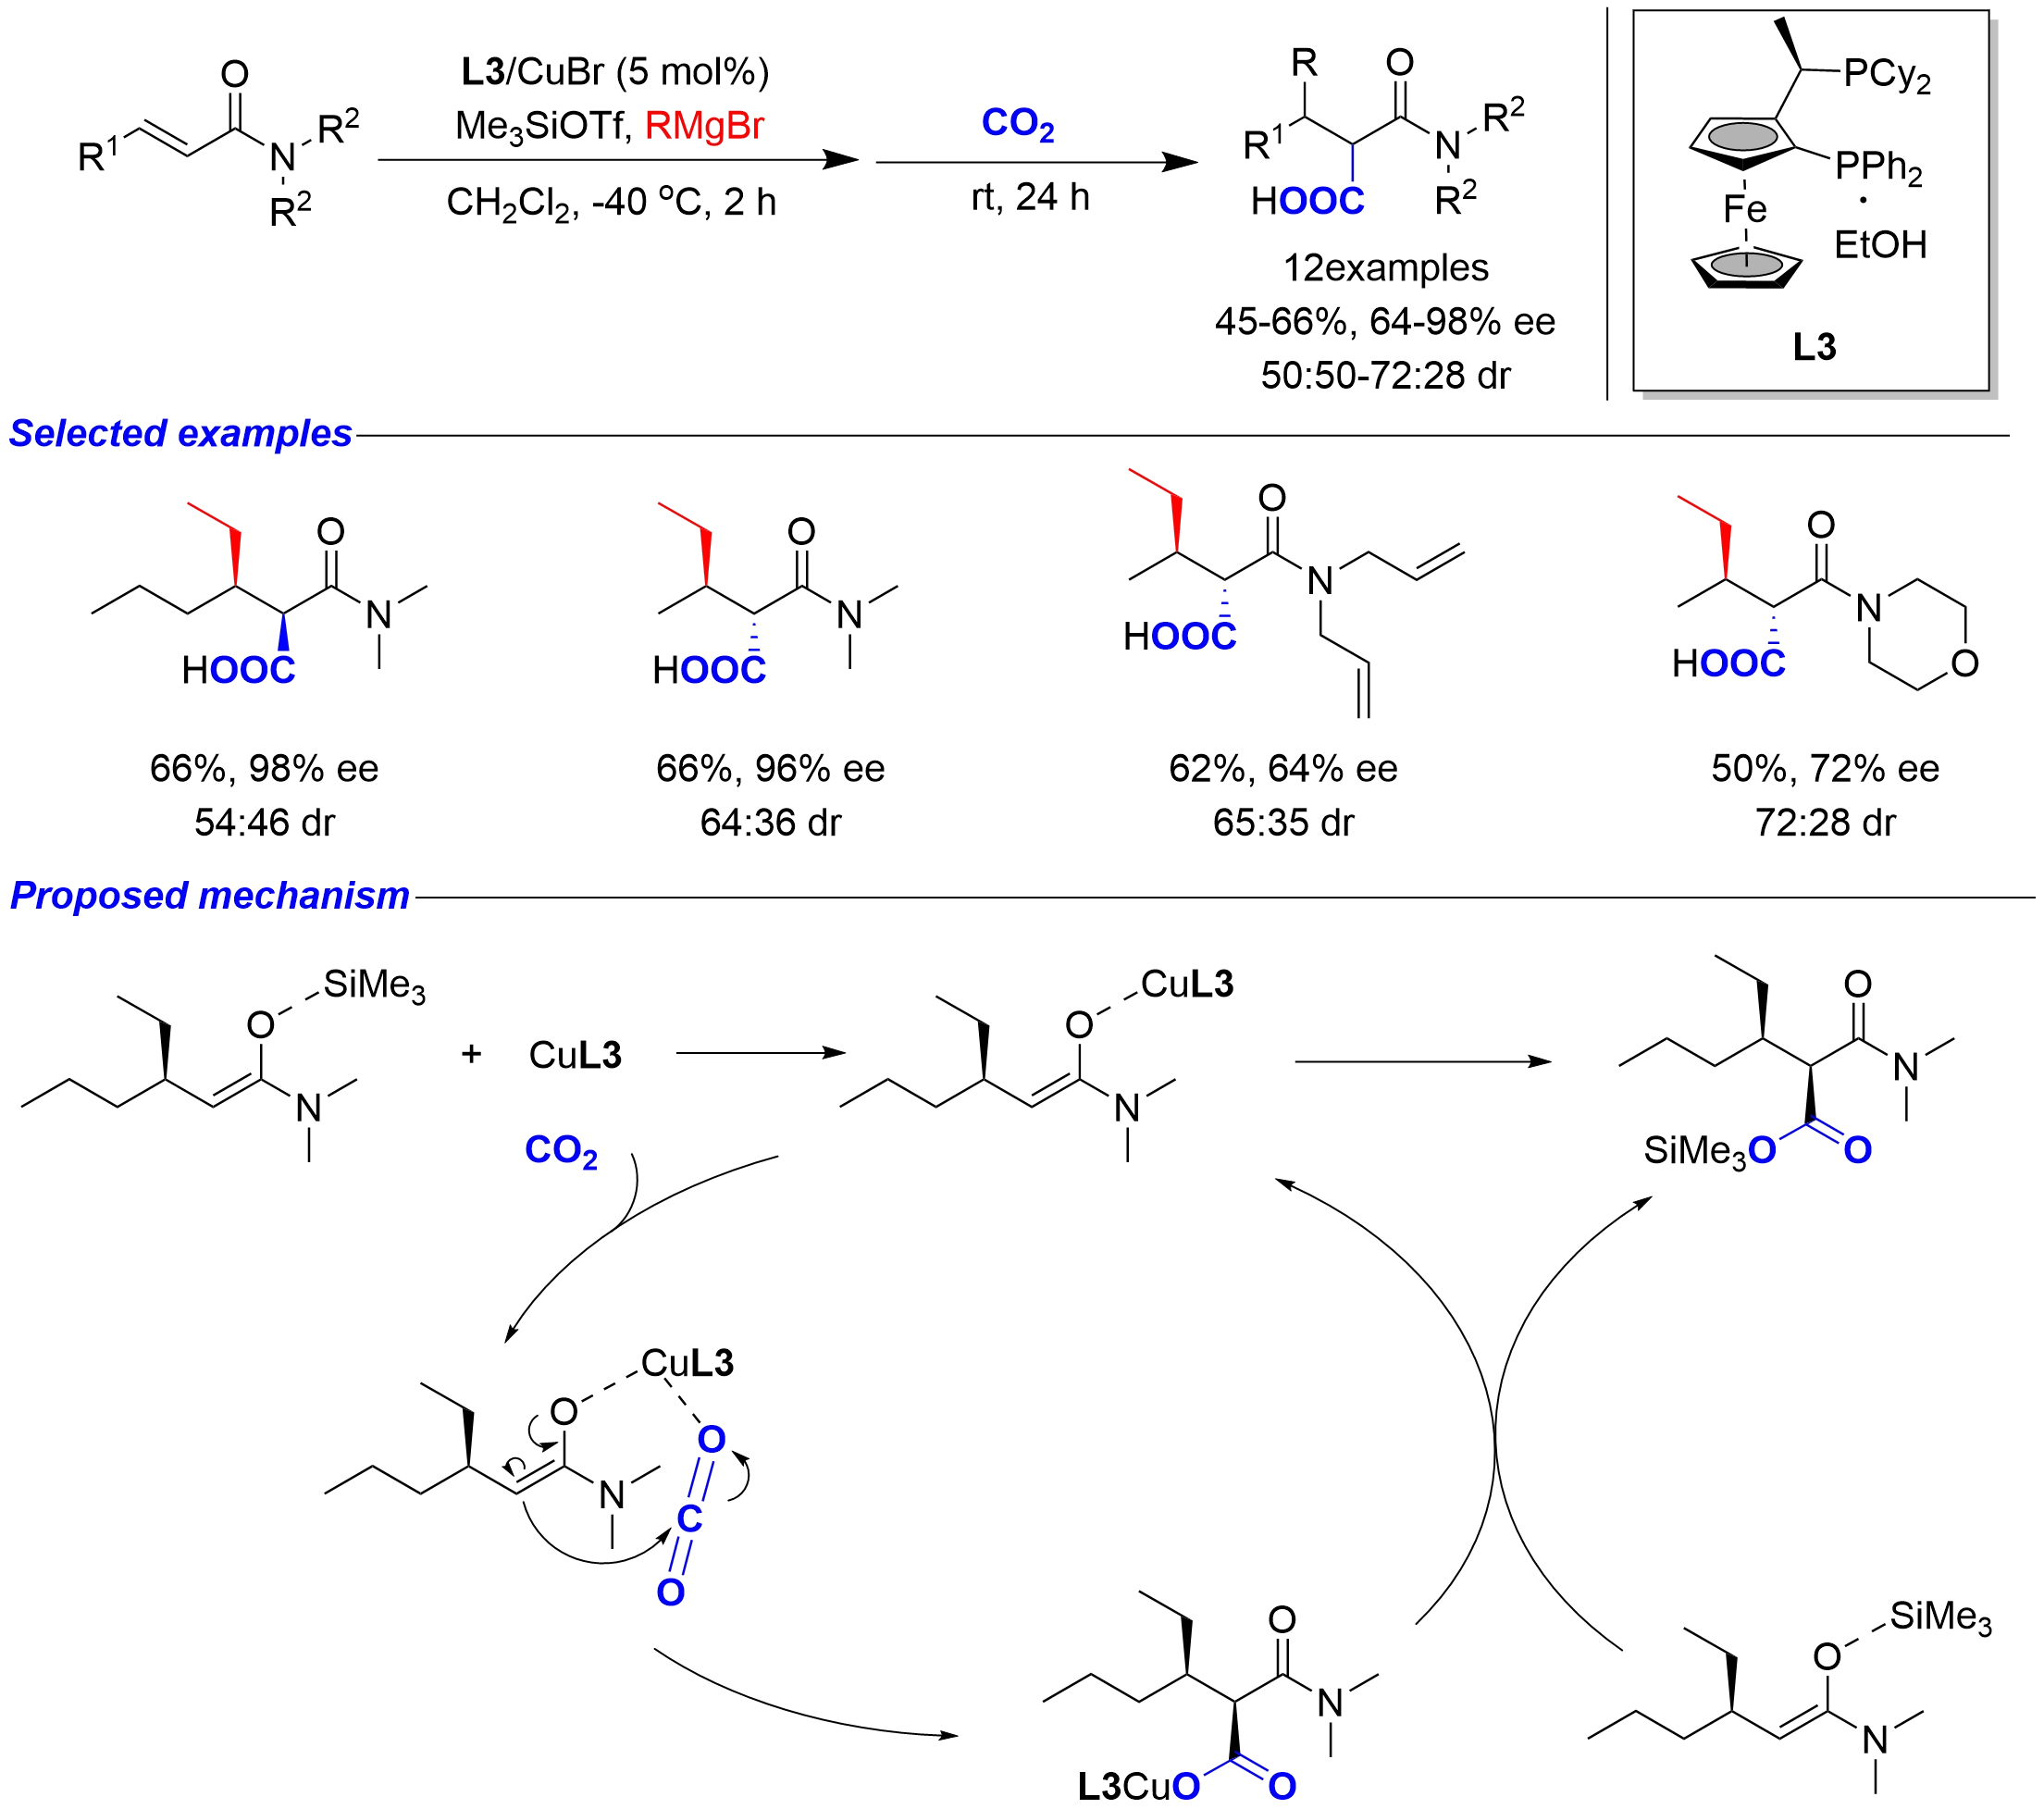
Scheme 5. Cu-catalyzed asymmetric conjugate addition/CO2 capture to chiral α-carboxyl amides.
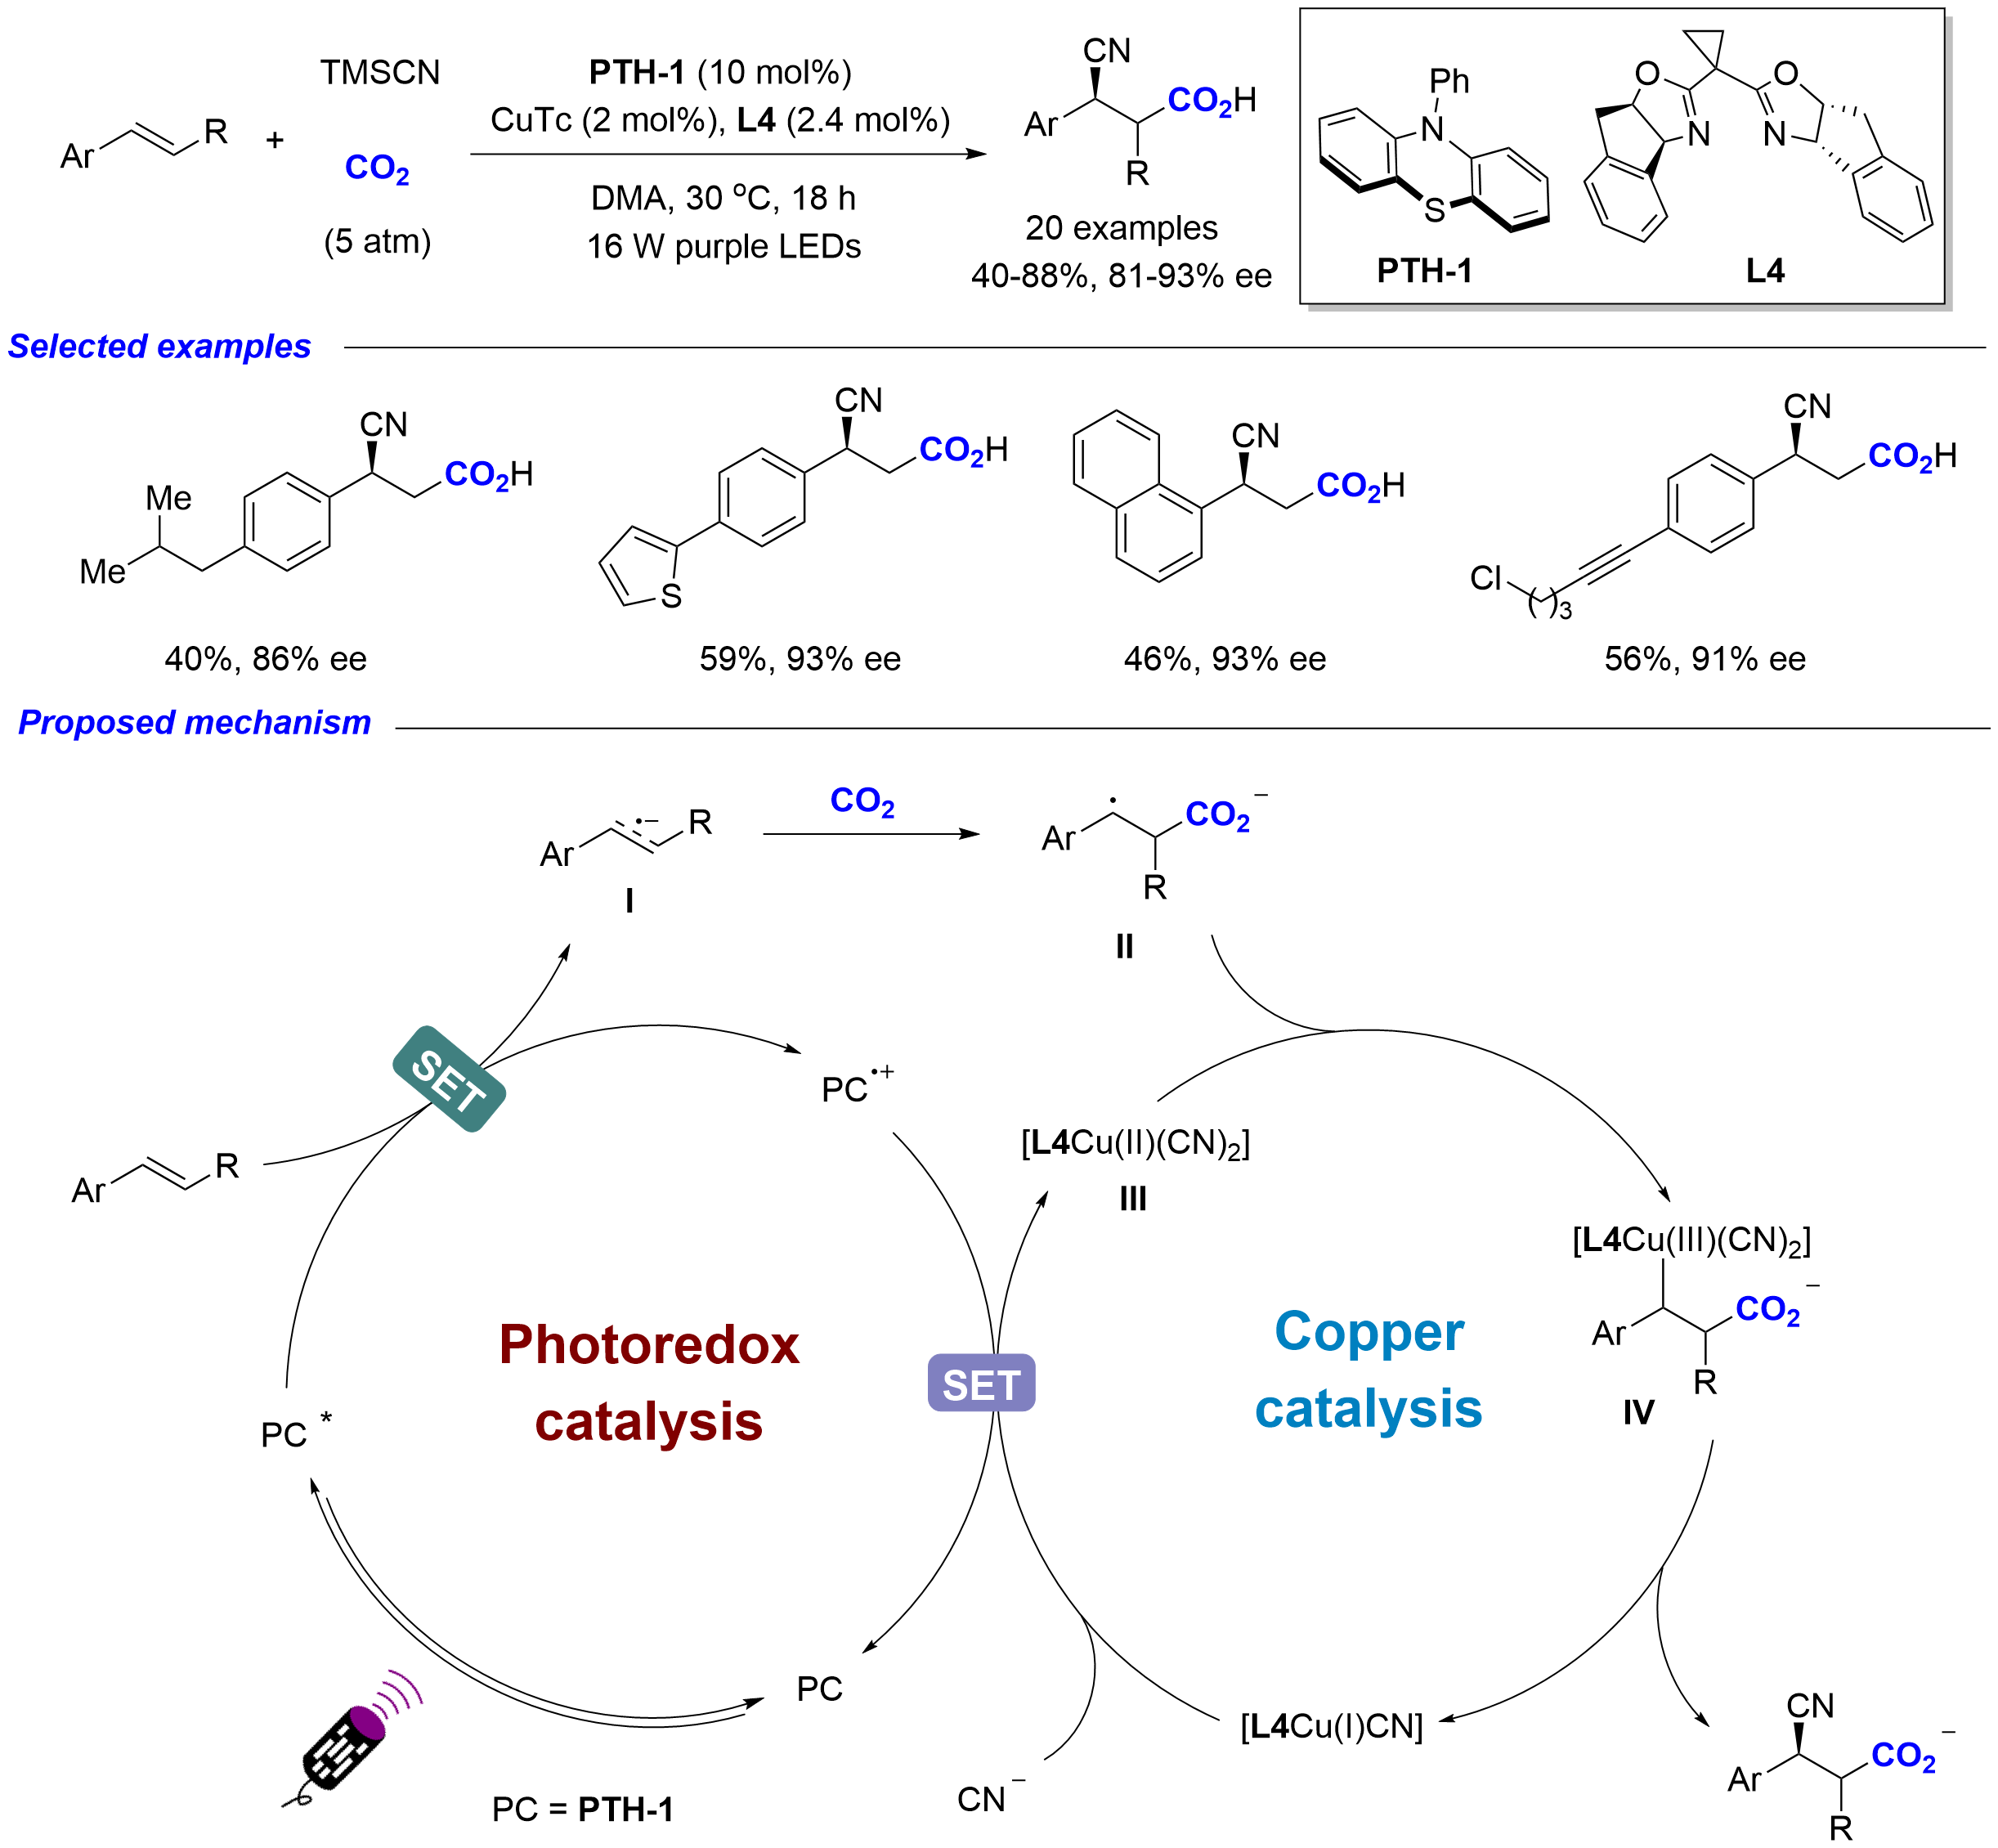
In 2024, Chen, Xiao and co-workers reported a photoredox/copper cooperative catalytic strategy for the cyanocarboxylation of aromatic alkenes, providing a range of α-chiral nitriles with high regio-, chemo-, and enantioselectivities (Scheme 6)[22]. With the combination of organic photocatalyst (PTH-1) and chiral copper catalyst under irradiation with purple LEDs, the desired chiral carboxylic acids were obtained in up to 88% yield and 93% ee. The substrate scope is broad, and various functional groups as well as complex drug-like scaffolds are tolerated. On the basis of mechanistic experiments and DFT calculations, the following catalytic cycle is proposed: the excited-state photocatalyst reduces the alkene via single-electron transfer (SET) to generate a radical anion I, which then reacts with CO2 to form a benzylic radical II. Meanwhile, the Cu(I) catalyst coordinates with TMSCN and is oxidized by the oxidized photocatalyst to give a Cu(II)-dicyano species III, which undergoes enantioselective radical recombination with the benzylic radical, affording Cu(III) intermediate IV. Subsequently, this Cu(III) intermediate undergoes reductive elimination to deliver the chiral product and regenerate the Cu(I) catalyst, thereby closing both catalytic cycles.
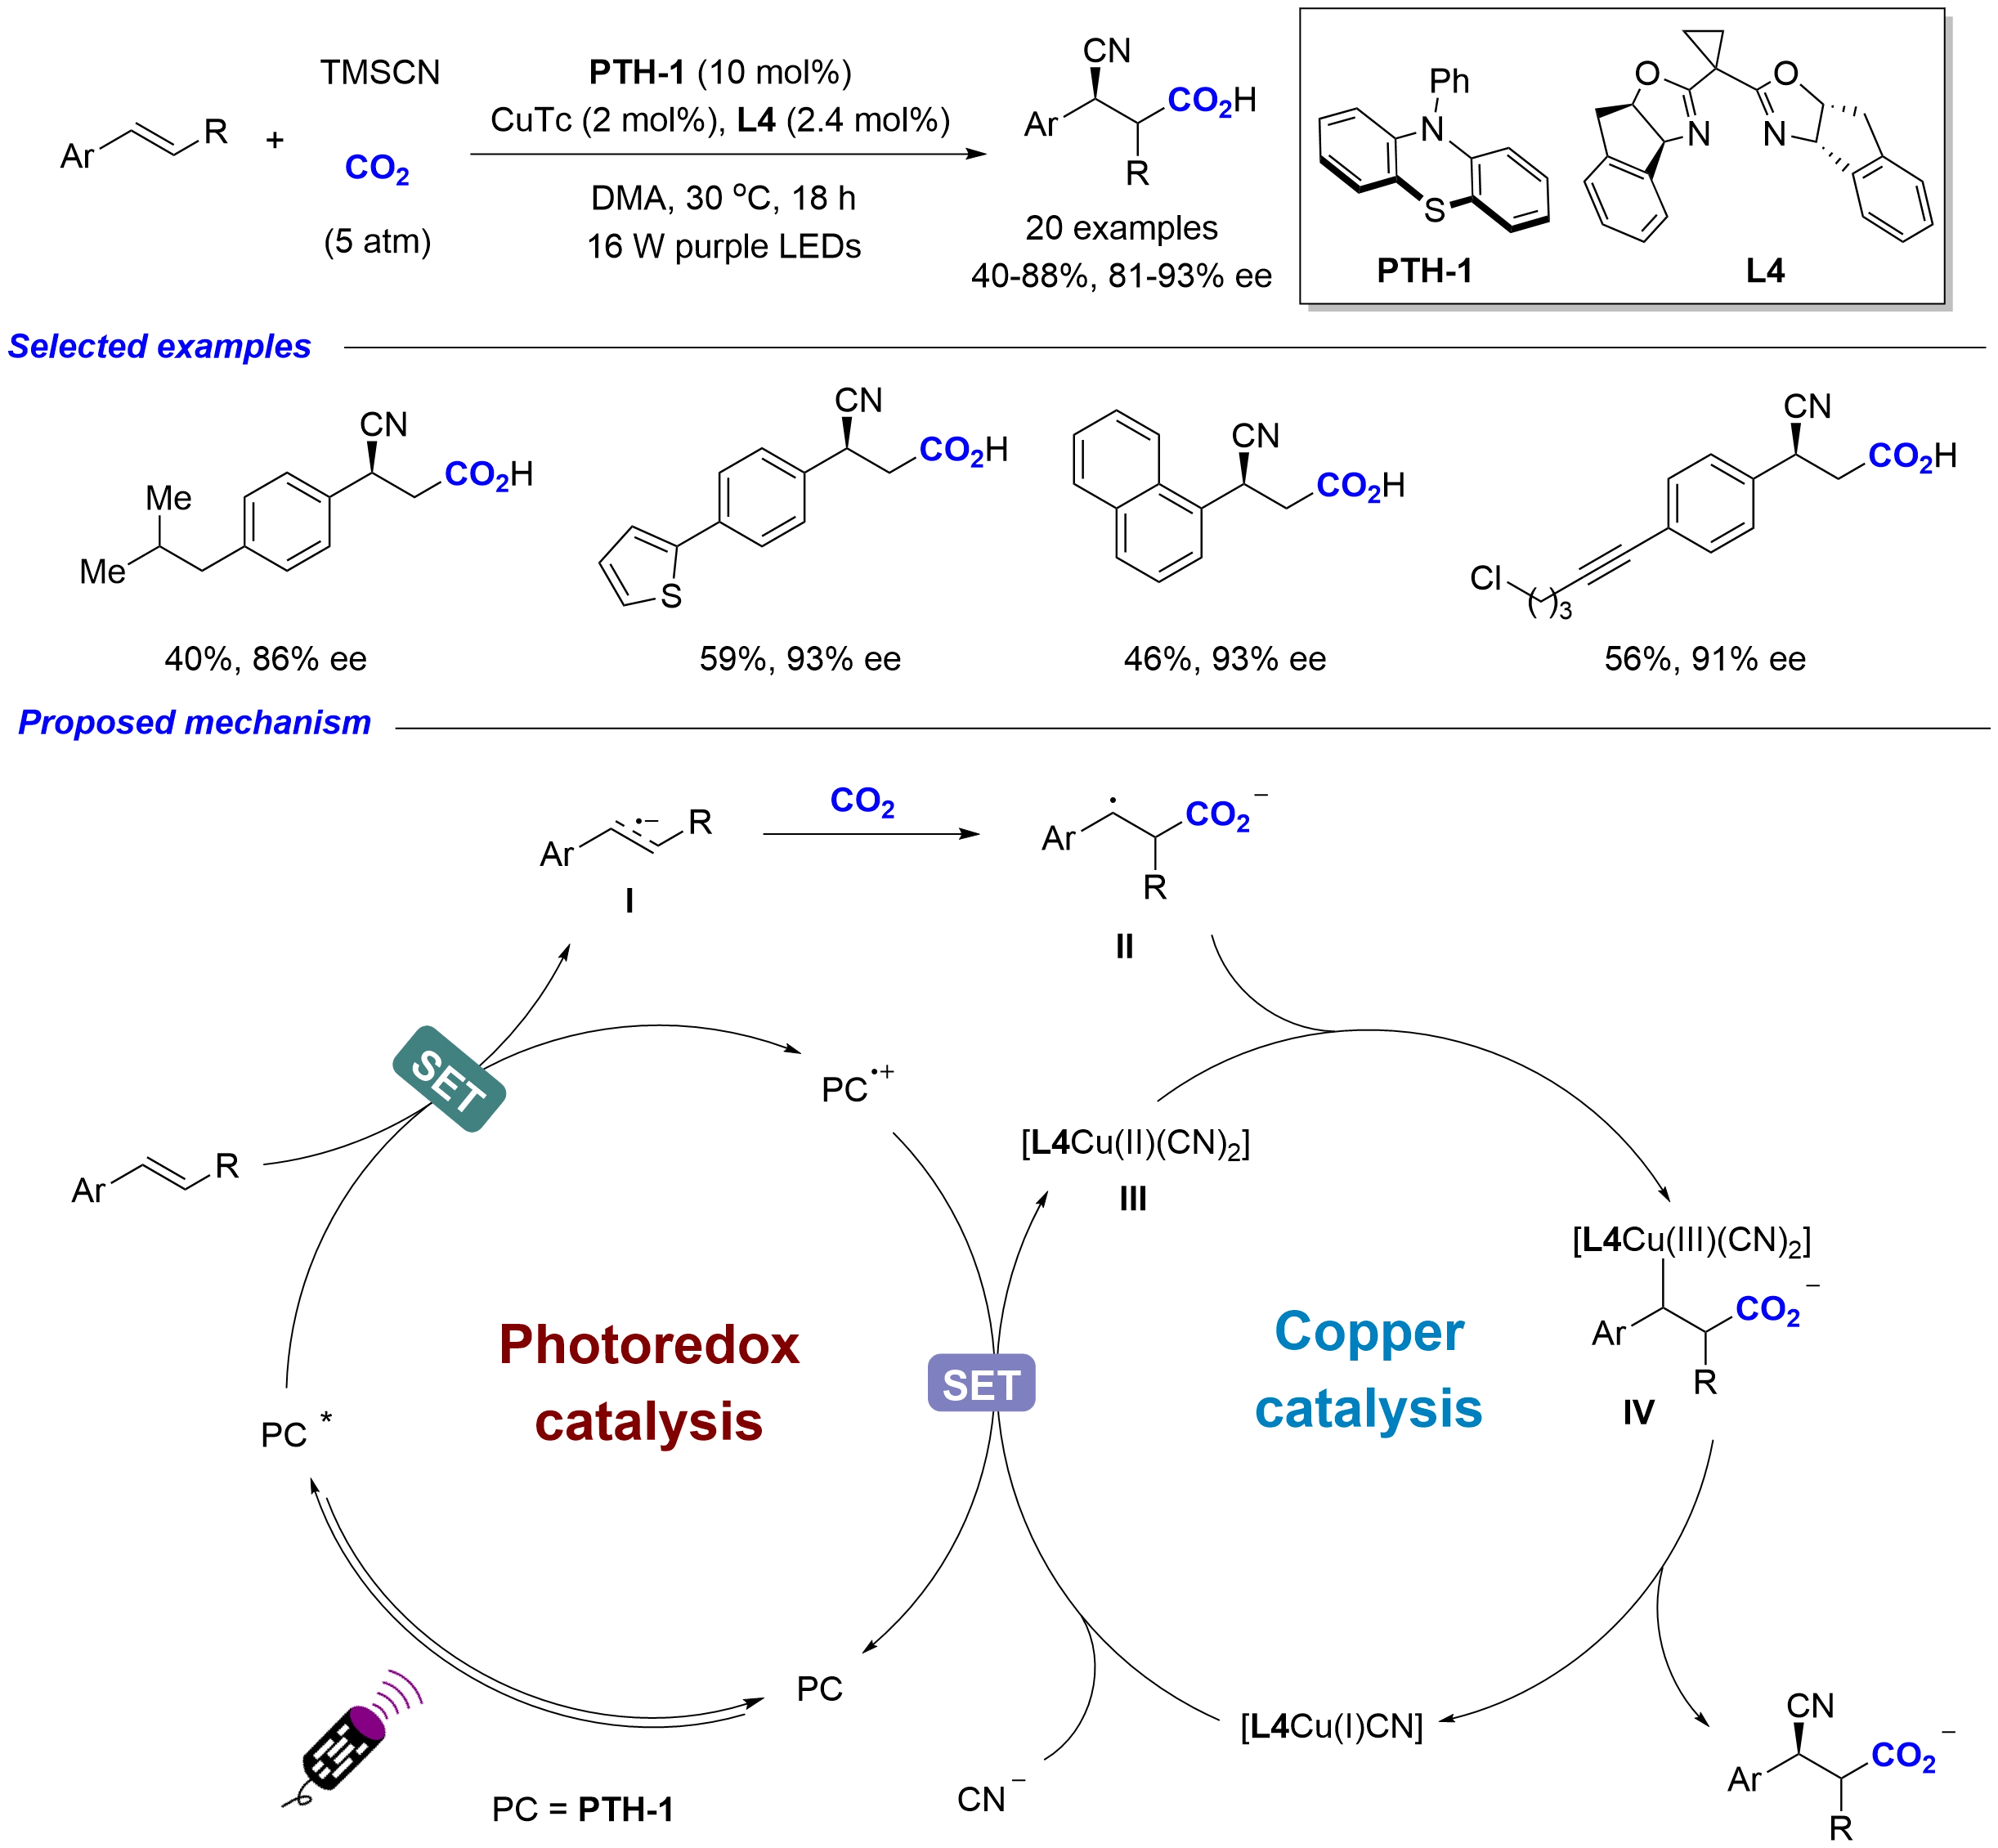
Scheme 6. Photoredox/copper cooperative catalysis for the cyanocarboxylation of aromatic alkenes.
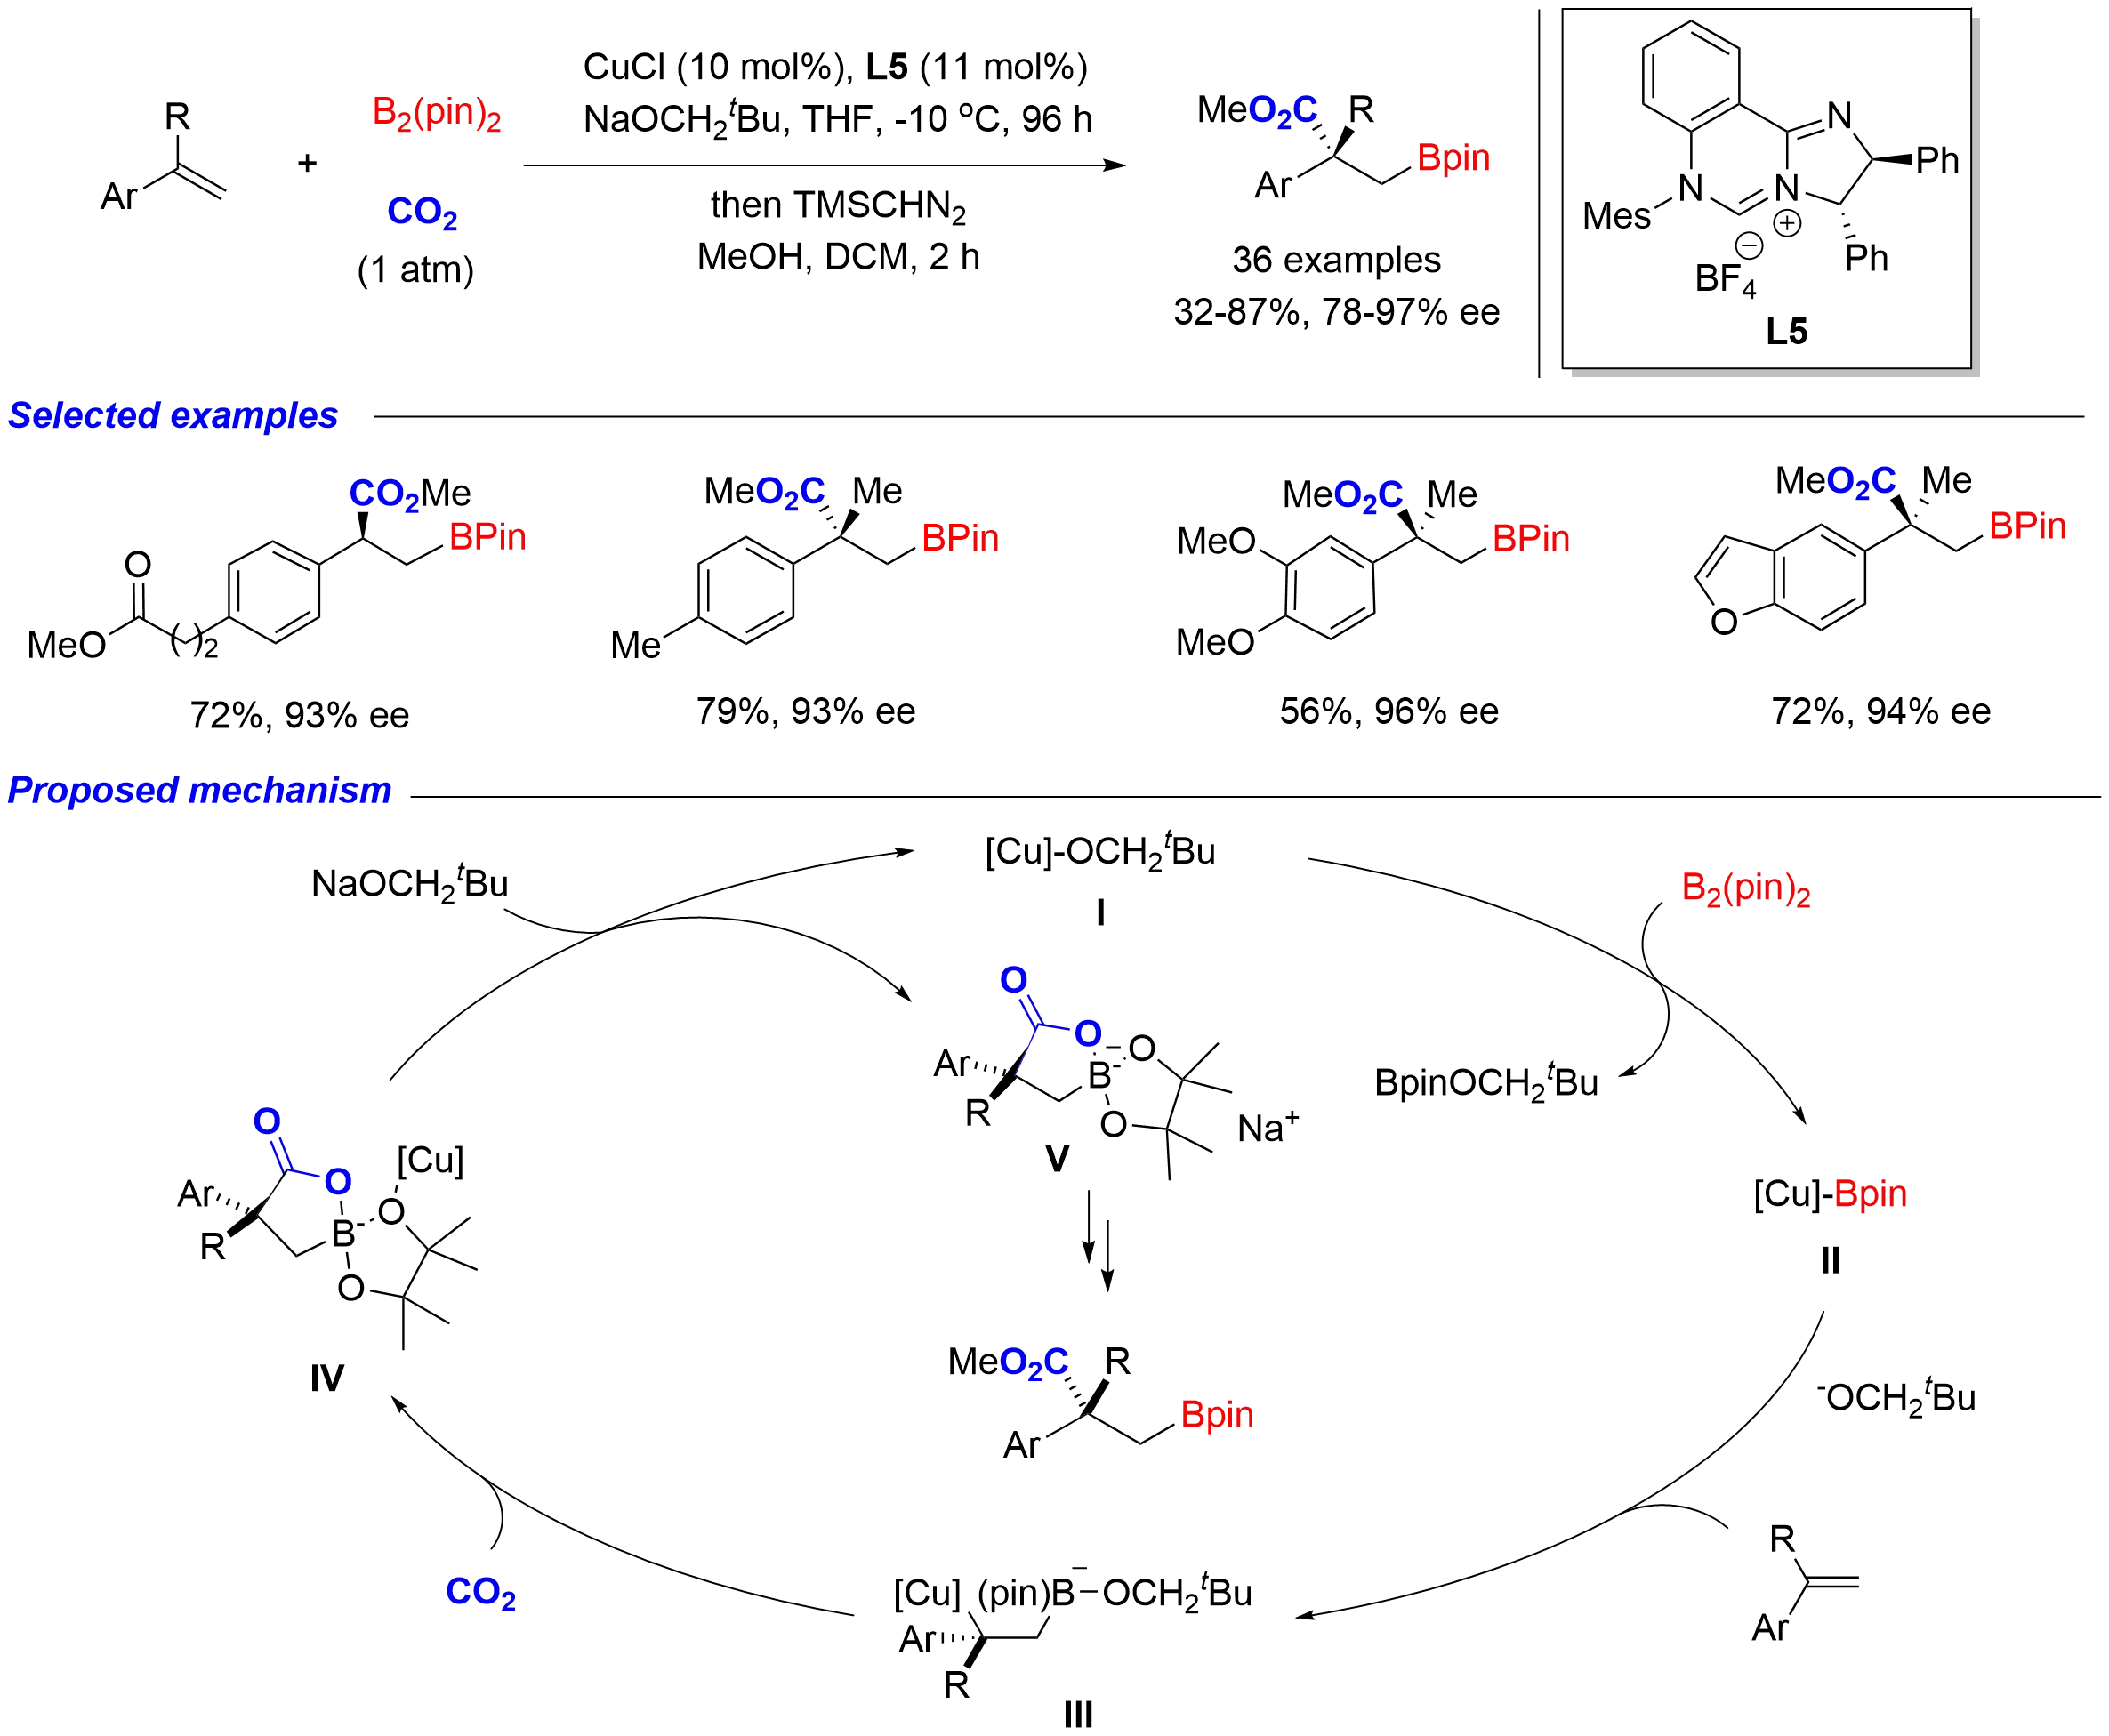
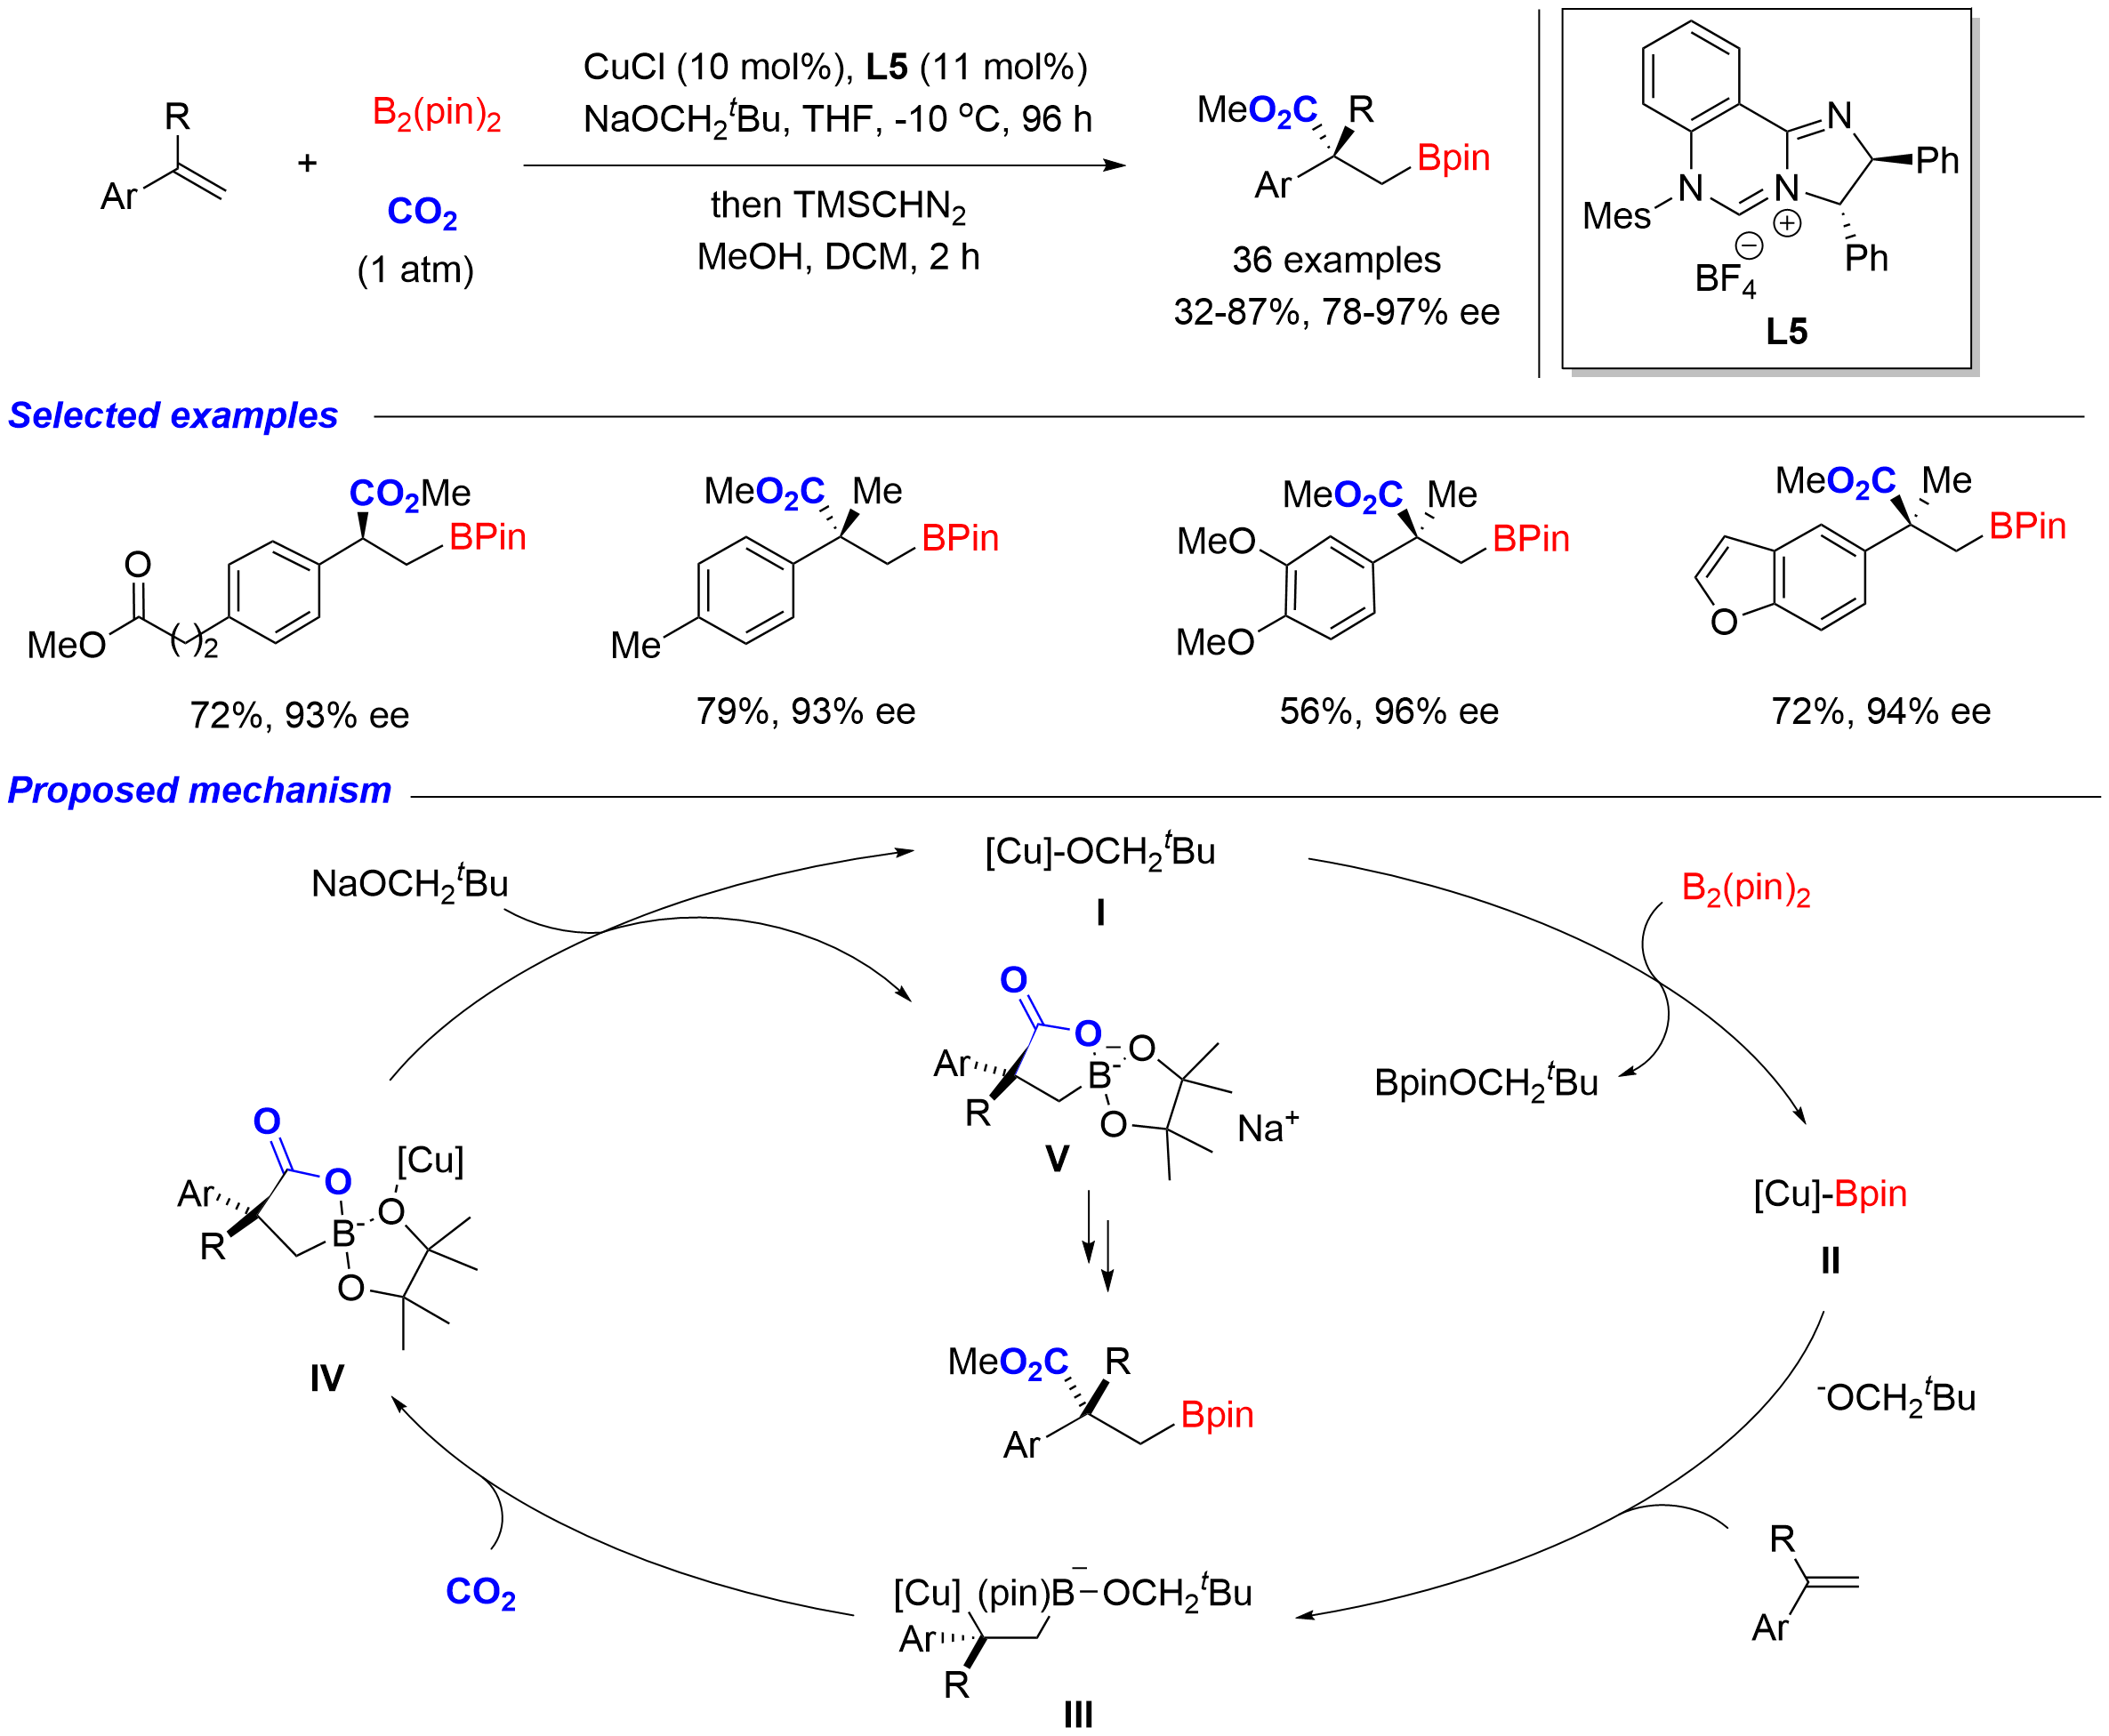
Boracarboxylation of alkenes. In 2024, Tang, Zhou and coworkers reported the first example of a Cu-catalyzed highly regio- and enantioselective boracarboxylation of arylalkenes with CO2 and bis(pinacolato)diboron [B2(pin)2] (Scheme 7)[23]. Using CuCl as the catalyst, a chiral NHC ligand, and NaOCH2tBu as the base, a variety of β-boron-functionalized α-carboxylic acids were obtained in up to 87% yield with up to 97% ee under mild conditions. It was proposed that the catalytic cycle proceeds as follows. Metathesis of NaOCH2tBu with the chiral NHC-CuCl complex generates intermediate I, which reacts with B2(pin)2 to afford the active species II. Species II undergoes Re-face boracupration of the alkene double bond, stereoselectively forming the key intermediate III. Subsequently, CO2 inserts directly into the Cu-benzyl bond in a stereoretentive manner to furnish the (R)-carboxylic acid intermediate IV. During this process, the alkoxy group bound to the B(pin) unit facilitates enantioselectivity control and accelerates the carboxylation step by generating vicinal steric hindrance and enhancing the nucleophilicity of the benzylic carbon. Finally, another equivalent of NaOCH2tBu promotes dissociation of the Cu-NHC complex to regenerate the active catalyst, while releasing the carboxylate product bearing a benzylic stereocenter.
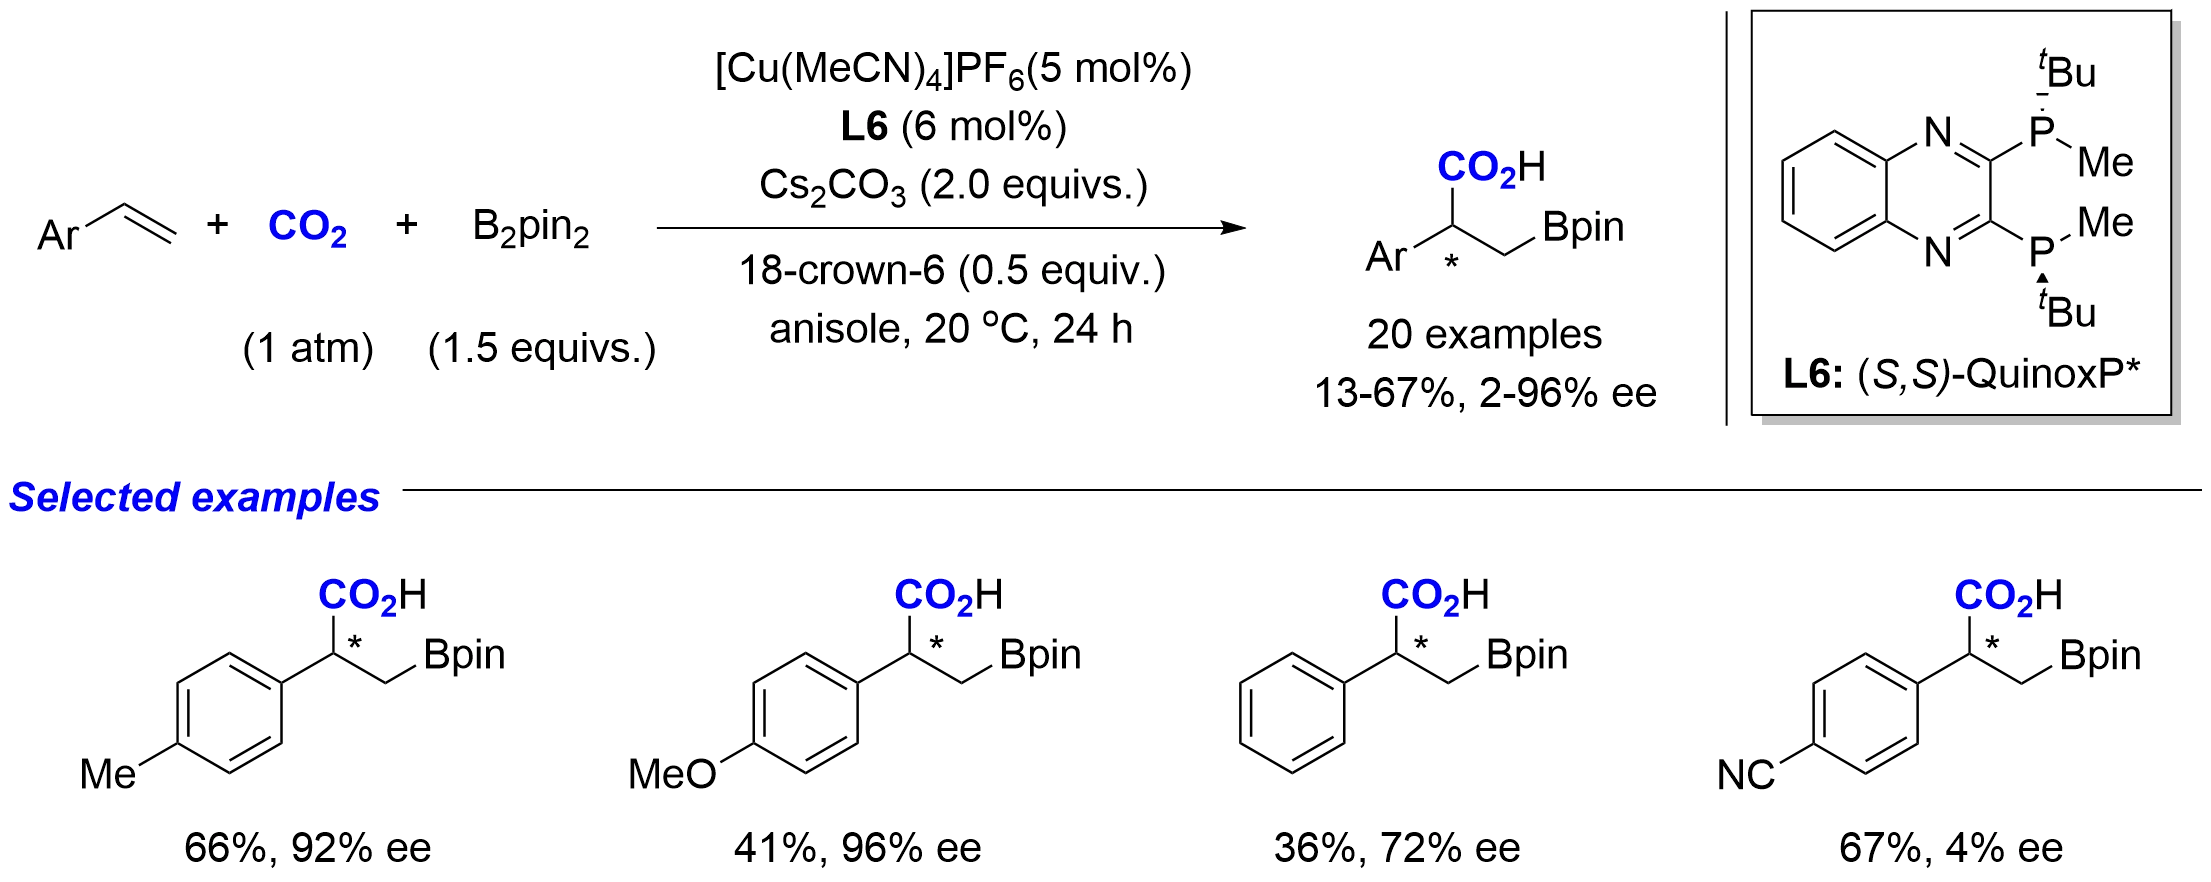
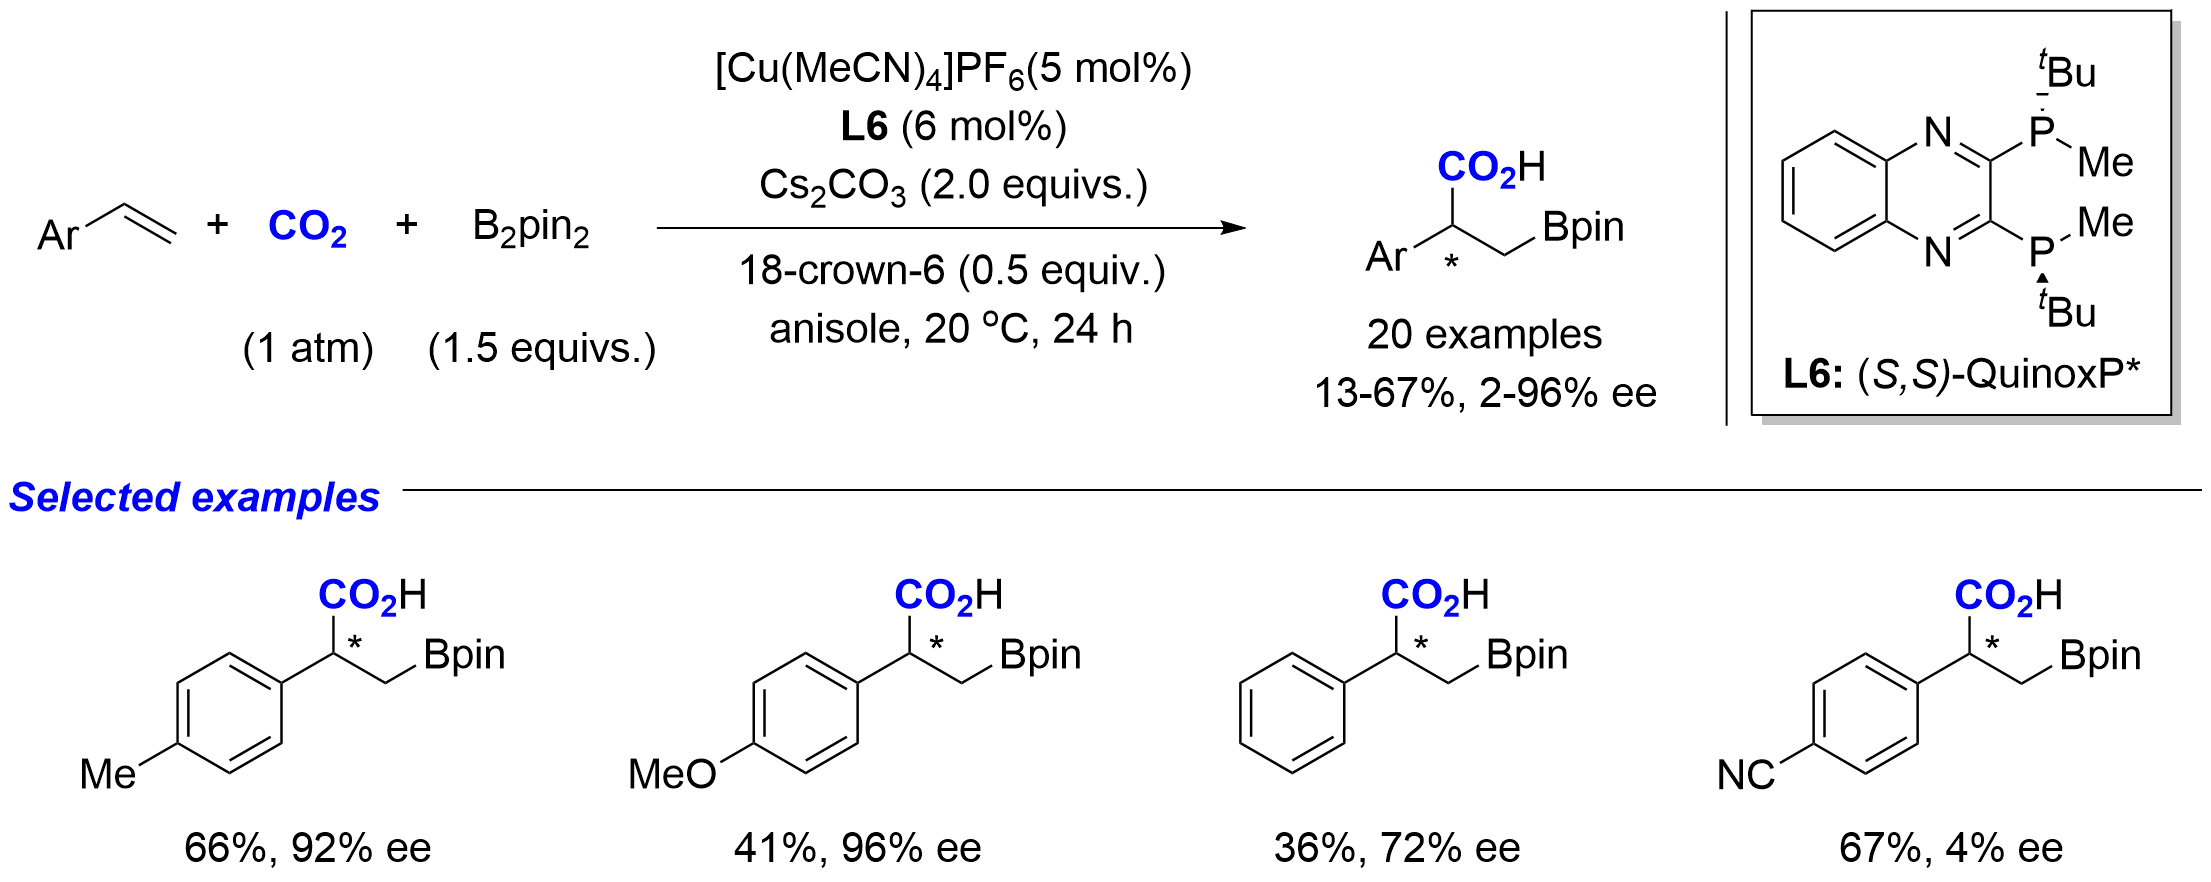
In the same year, Bayer, Hopmann and coworkers independently reported this reaction using a different catalytic system (Scheme 8)[24]. With [Cu(MeCN)4]PF6 as the copper source, (S, S)-QuinoxP* as the chiral ligand, Cs2CO3 as the base, and 18-crown-6 as an additive, the β-boron α-carboxylic acids were obtained in 13-67% yield with up to 96% ee. The electronic properties of the styrene ring significantly affected enantioselectivity: electron-rich substrates gave excellent selectivity, whereas electron-deficient substrates afforded low enantiomeric ratios. DFT calculations revealed that the regioselective boracupration of styrene and subsequent competition between inner- and outer-sphere CO2 insertion pathways govern the enantioselectivity.
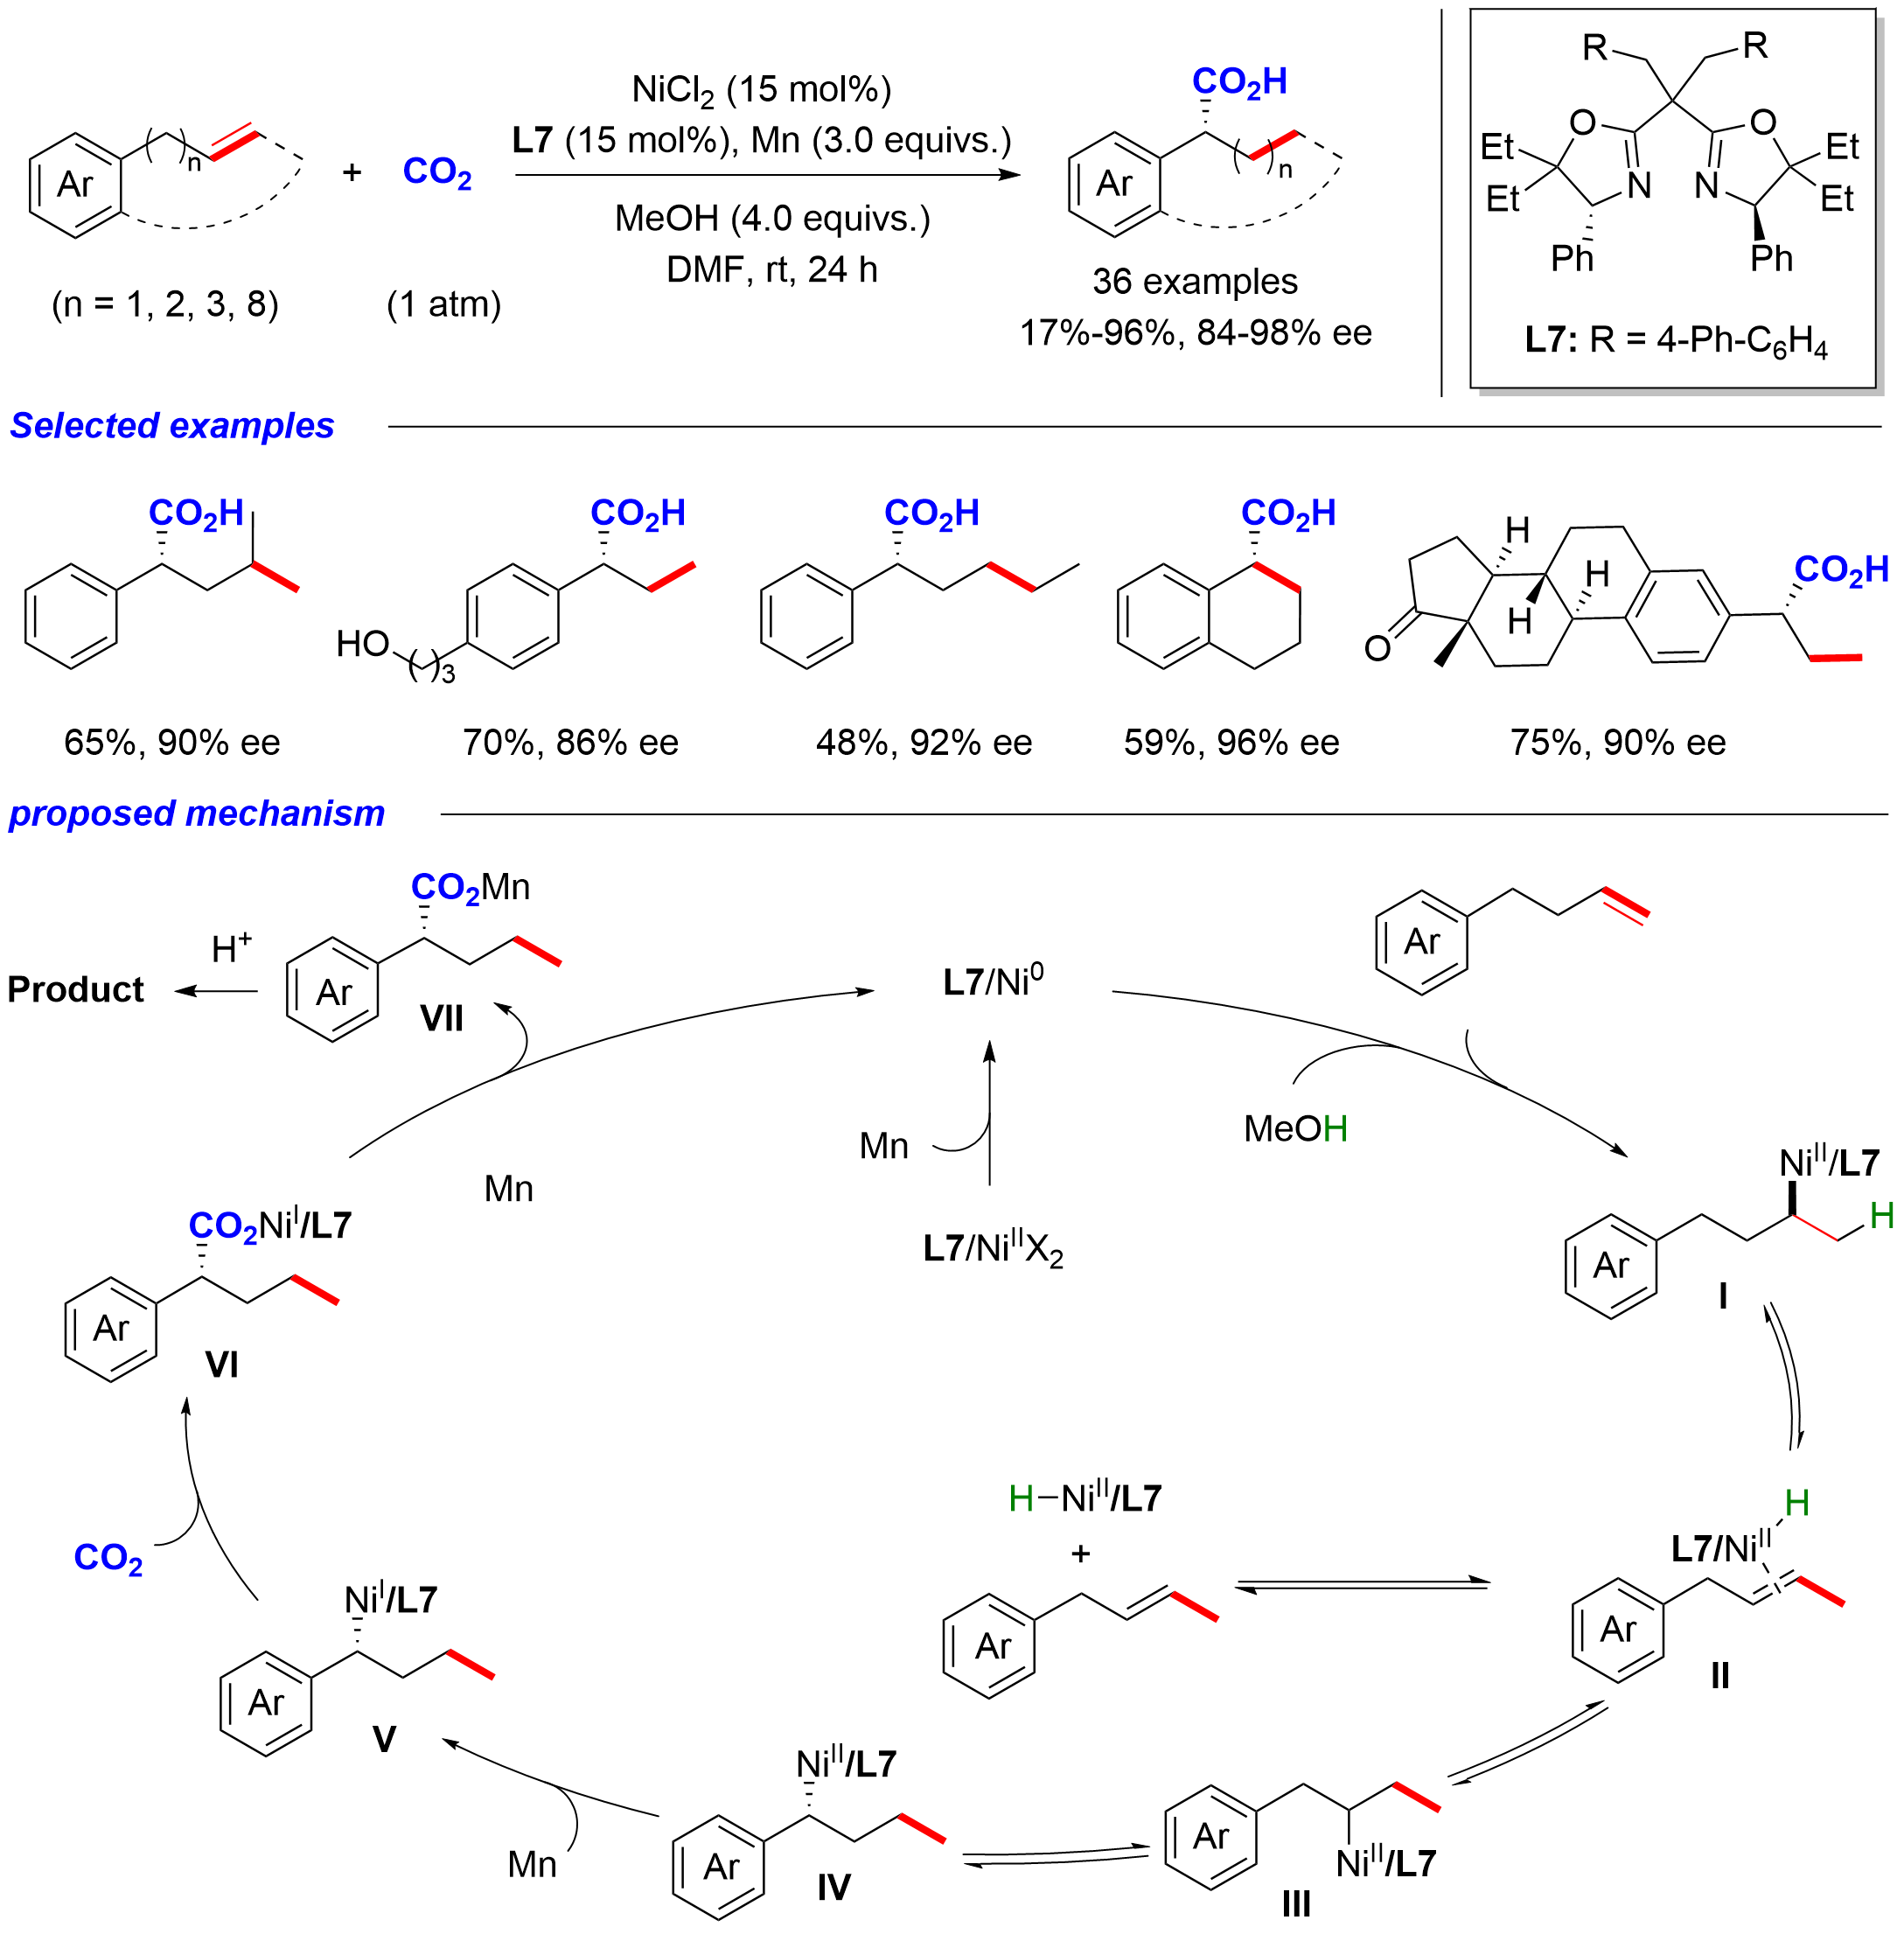
Hydrocarboxylation of alkenes. Apart from the enantioselective boracarboxylation approach, the hydrocarboxylation of alkenes with CO2 represents another important strategy for constructing chiral carboxylic acids. In 2024, Tang, Zhou and coworkers reported an enantioselective nickel-catalyzed migratory hydrocarboxylation of unactivated alkenes with CO2 (Scheme 9)[25]. This strategy employed a chiral side-arm bisoxazoline (SaBOX) ligand to achieve chain-walking under mild conditions, converting remote alkenes into thermodynamically stable α-chiral benzylic nickel intermediates. Subsequent CO2 insertion and protonation afforded α-chiral carboxylic acids with excellent regio- and enantioselectivities. Unactivated terminal alkenes, internal alkenes, and benzo-fused cycloalkenes were all viable substrates, delivering the desired α-chiral carboxylic acids in up to 96% yield with up to 98% ee. Moreover, an economical two-step synthesis of the antiplatelet aggregation drug (R)-indobufen from commercially available alkene and CO2 was also realized.
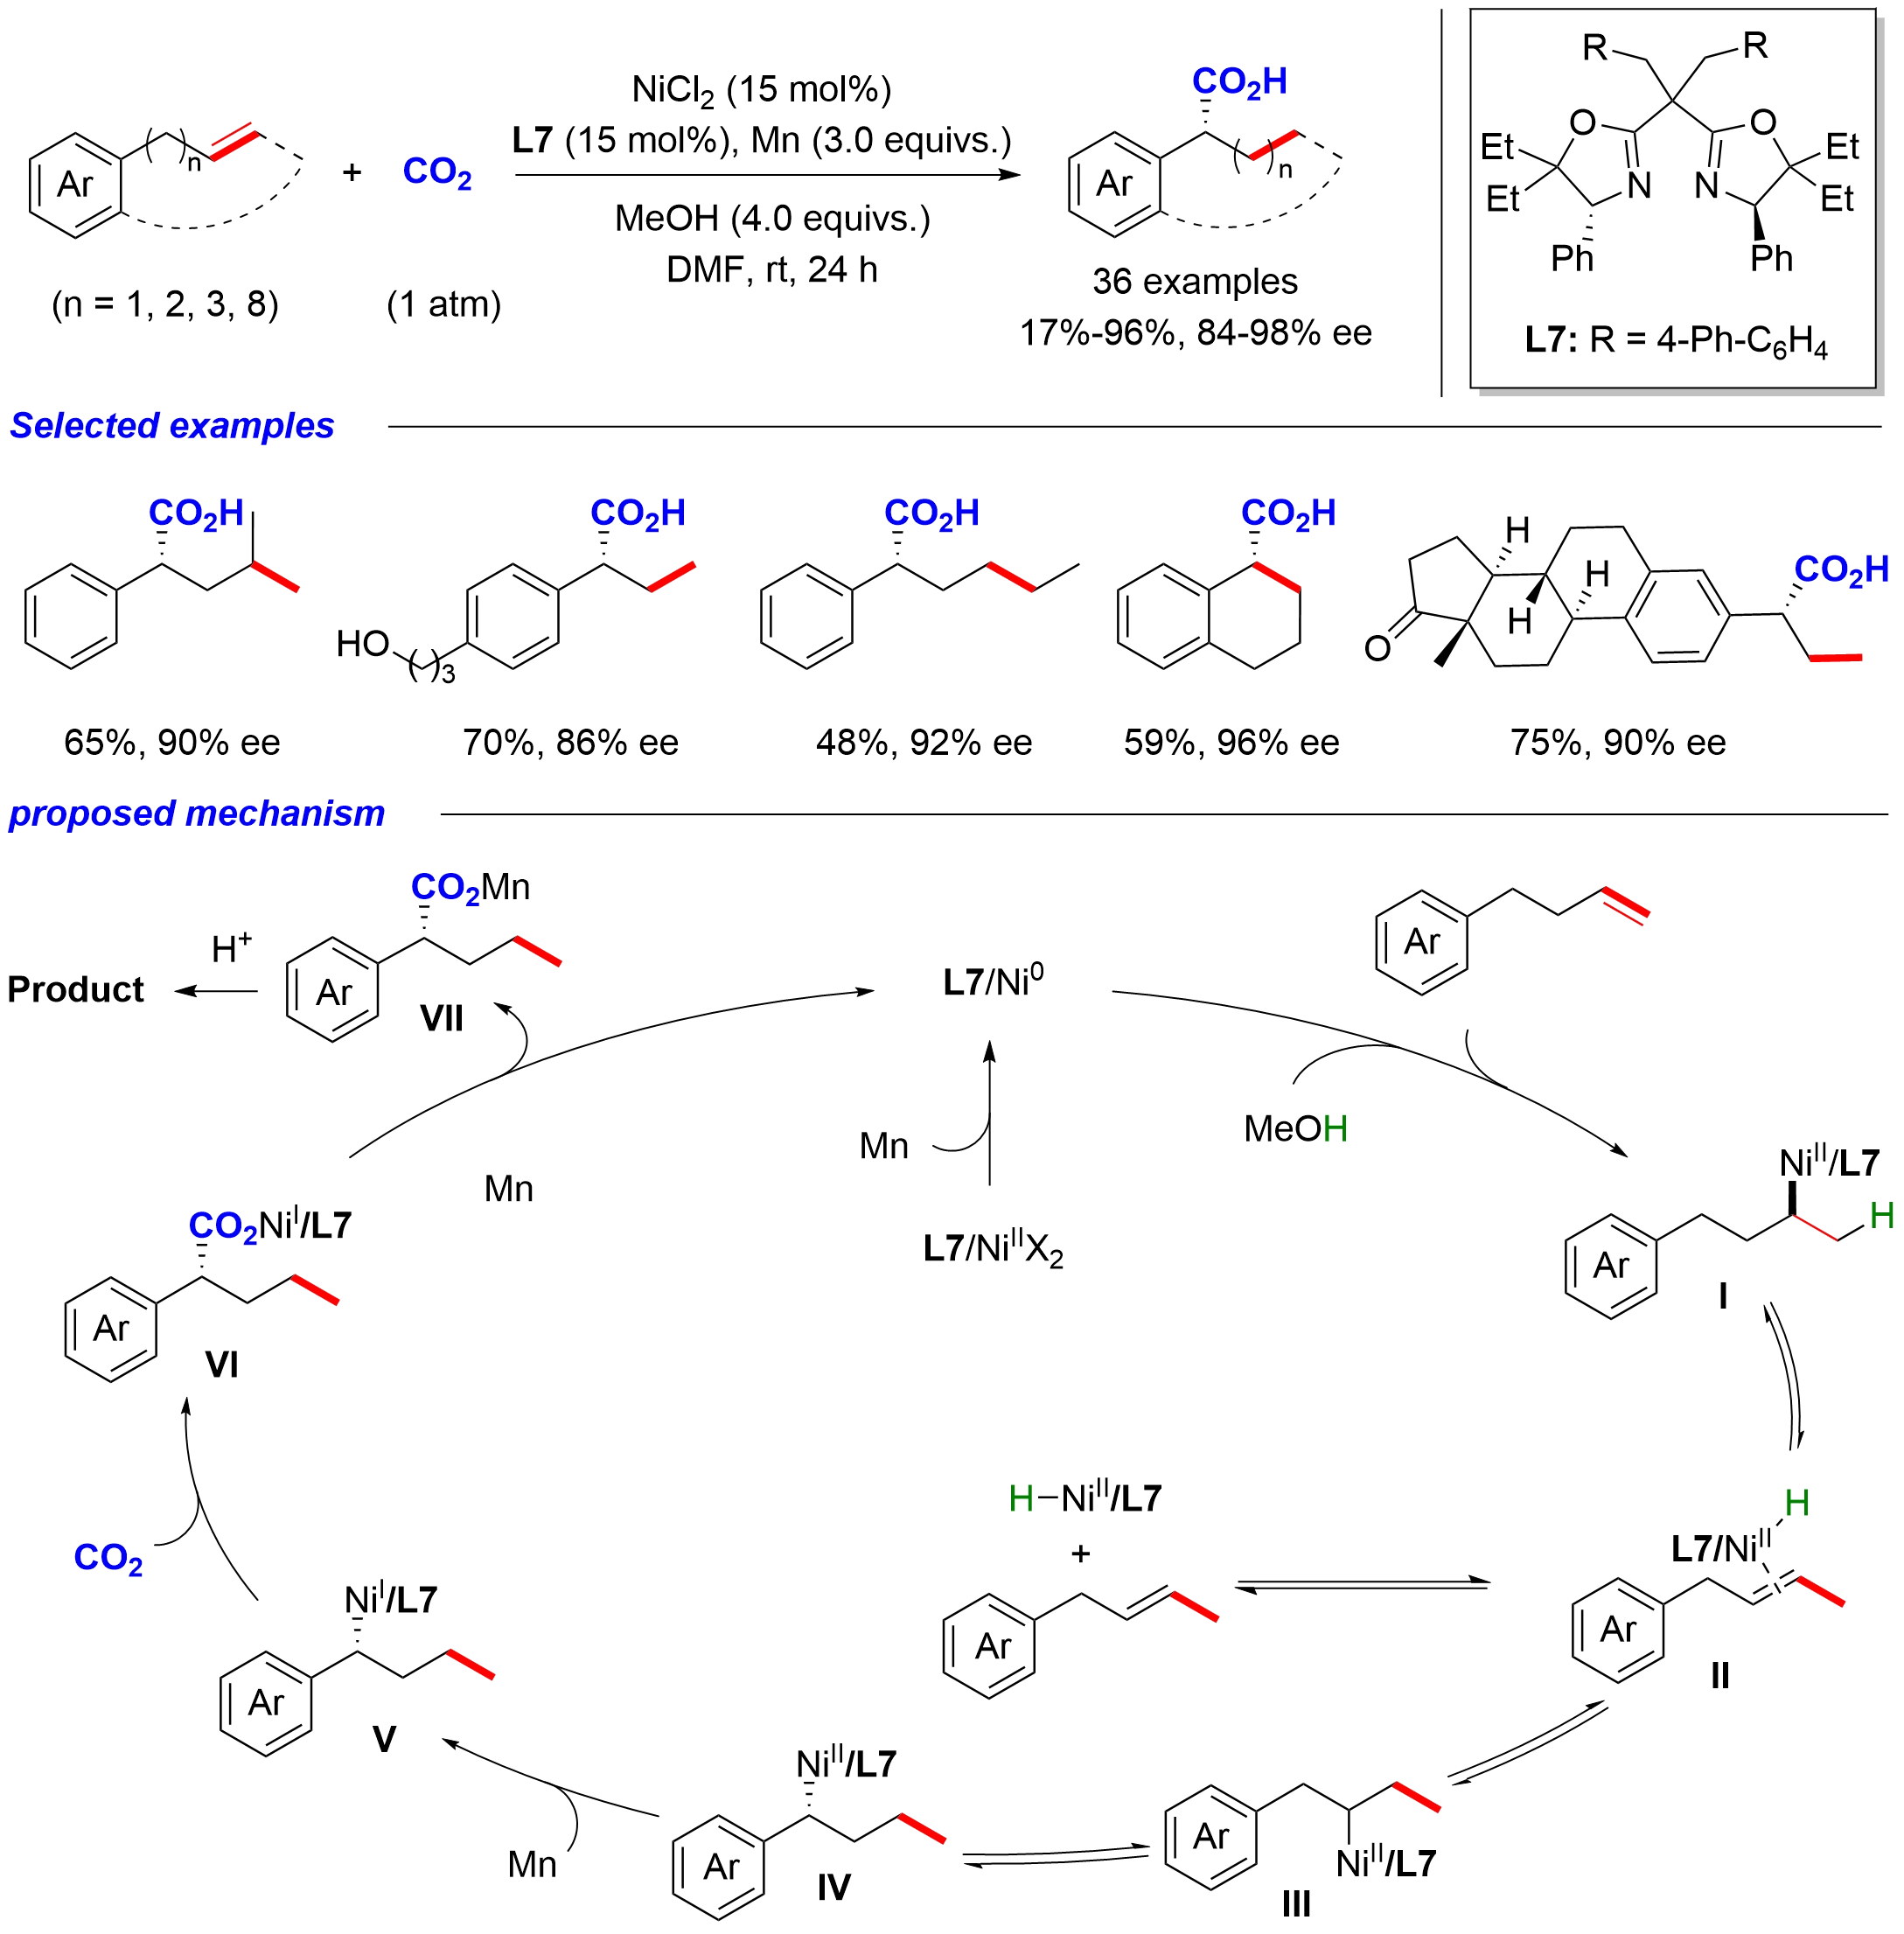
Deuterium labeling experiments confirmed that MeOH serves as the hydrogen source and that reversible dissociation of the alkene-nickel hydride complex occurs during chain-walking. The catalytic cycle proceeds as follows. The in situ generated SaBOX/Ni(II) precursor is reduced by Mn powder to afford the active L7/Ni(0) species. Subsequently, L7/Ni(0) undergoes oxidative addition with the O-H bond of methanol to generate a L7/Ni(II)-H species. This hydride species undergoes migratory insertion into the alkene, furnishing the Ni(II)-alkyl intermediate I. Under the control of the SaBOX ligand, the reaction favors a chain-walking process, wherein intermediate I undergoes a rapid and reversible iterative sequence of β-hydride elimination and migratory insertion, ultimately affording the thermodynamically stable benzylic nickel intermediate IV with high regio- and enantioselectivity. During this process, the alkene can reversibly dissociate from the nickel complex. Reduction of IV by Mn powder generates the α-chiral benzylic Ni(I) intermediate V, which subsequently undergoes insertion with CO2 to afford the chiral α-aryl carboxylic acid nickel species VI. Finally, VI is further reduced by Mn to regenerate the L7/Ni(0) species and produce the manganese carboxylate VII, which is converted into the target carboxylic acid product upon protonation during workup.
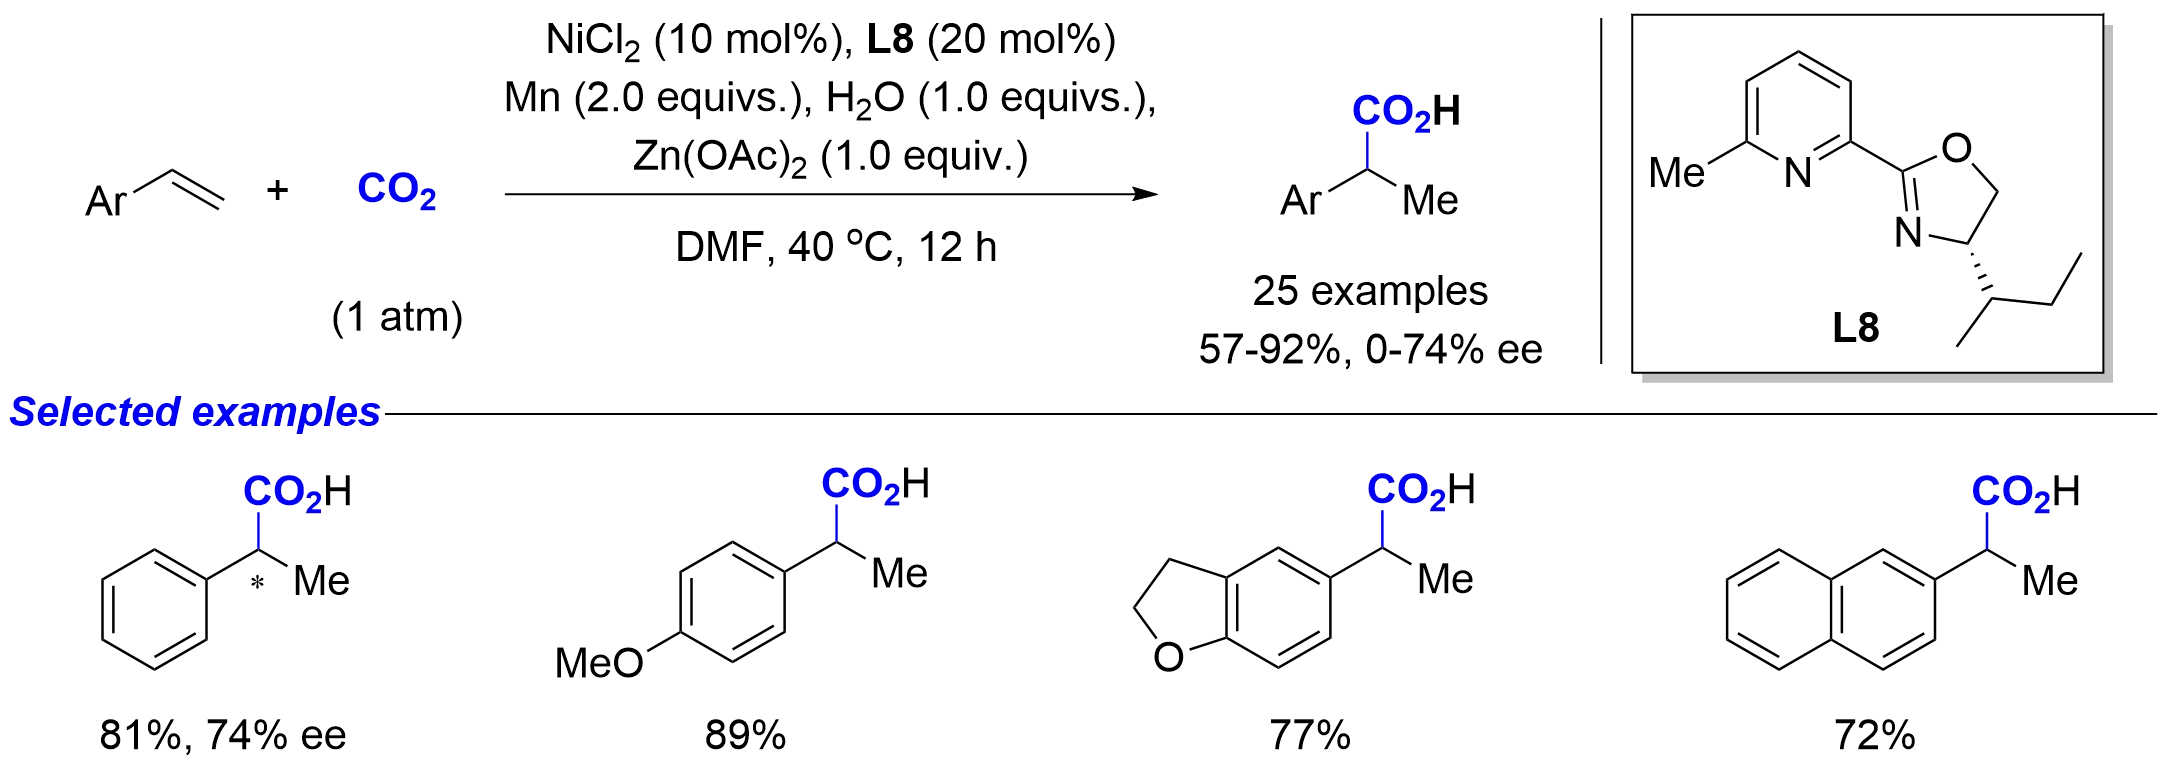
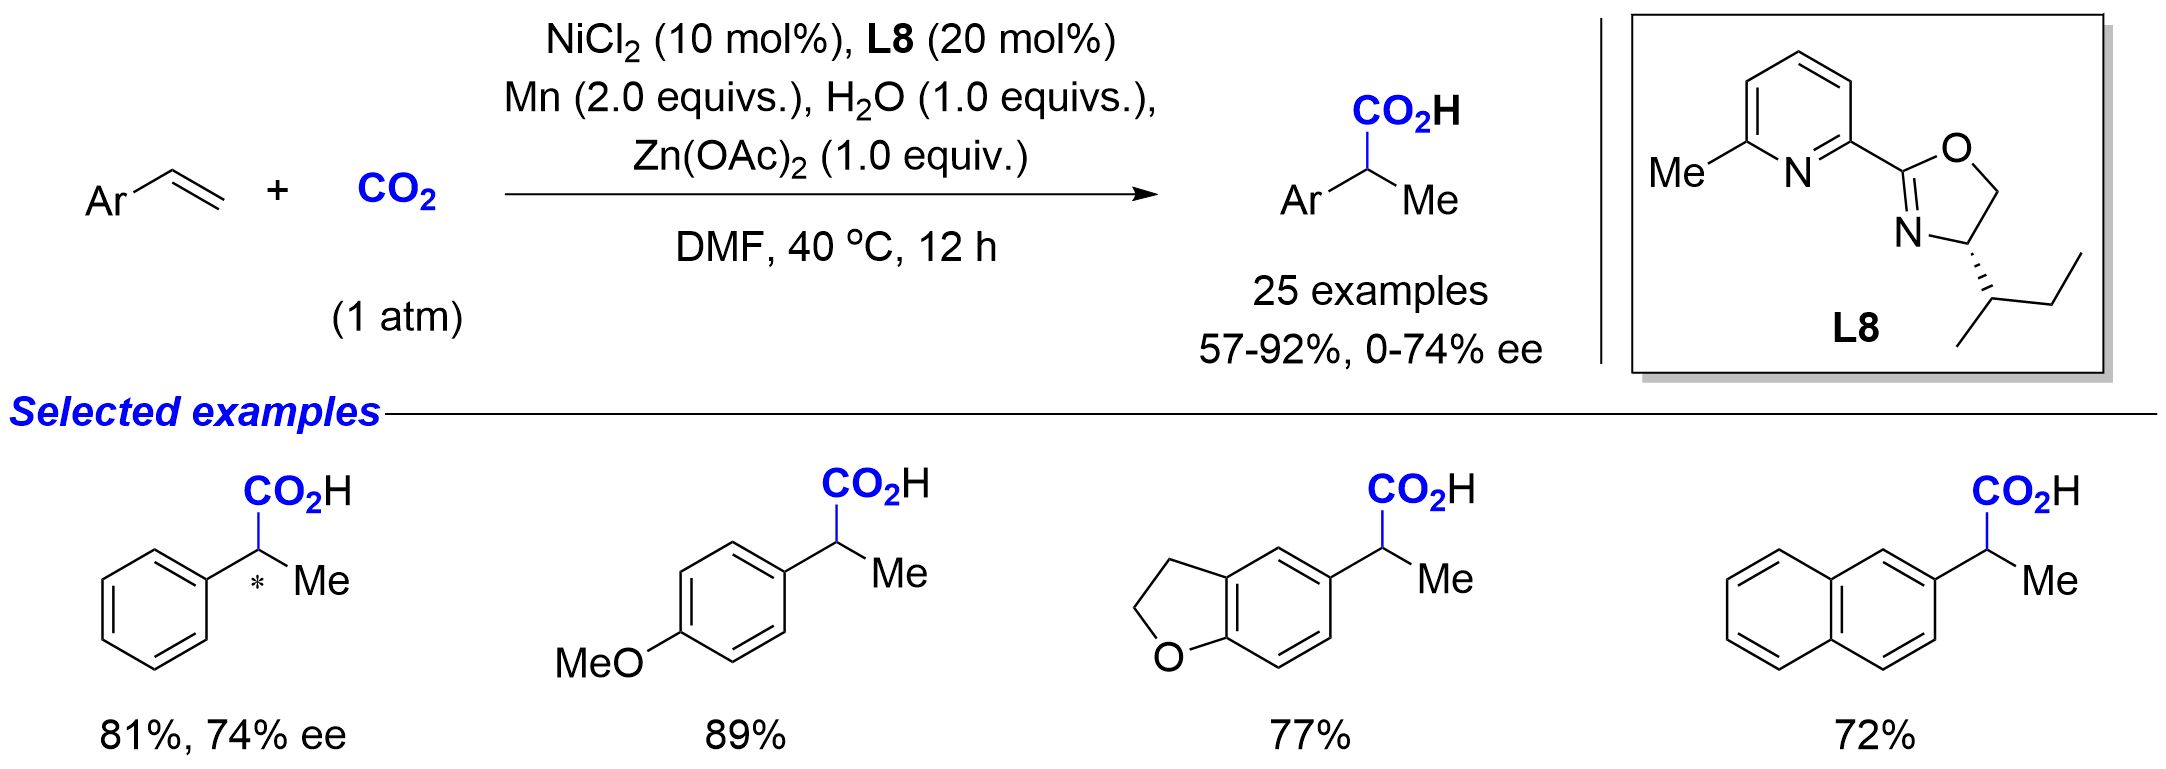
Subsequently, in 2025, Chen and coworkers reported a direct hydrocarboxylation strategy without chain-walking. Using water as the hydride source, Mn powder as the reducing agent, and zinc acetate as a key additive, the regioselective hydrocarboxylation of styrenes with CO2 was achieved under nickel catalysis (Scheme 10)[26]. Asymmetric catalysis was preliminarily realized using a chiral pyridine-oxazoline ligand, and the reaction of styrene with CO2 delivered the corresponding chiral α-phenyl carboxylic acid in 81% yield with 74% ee.
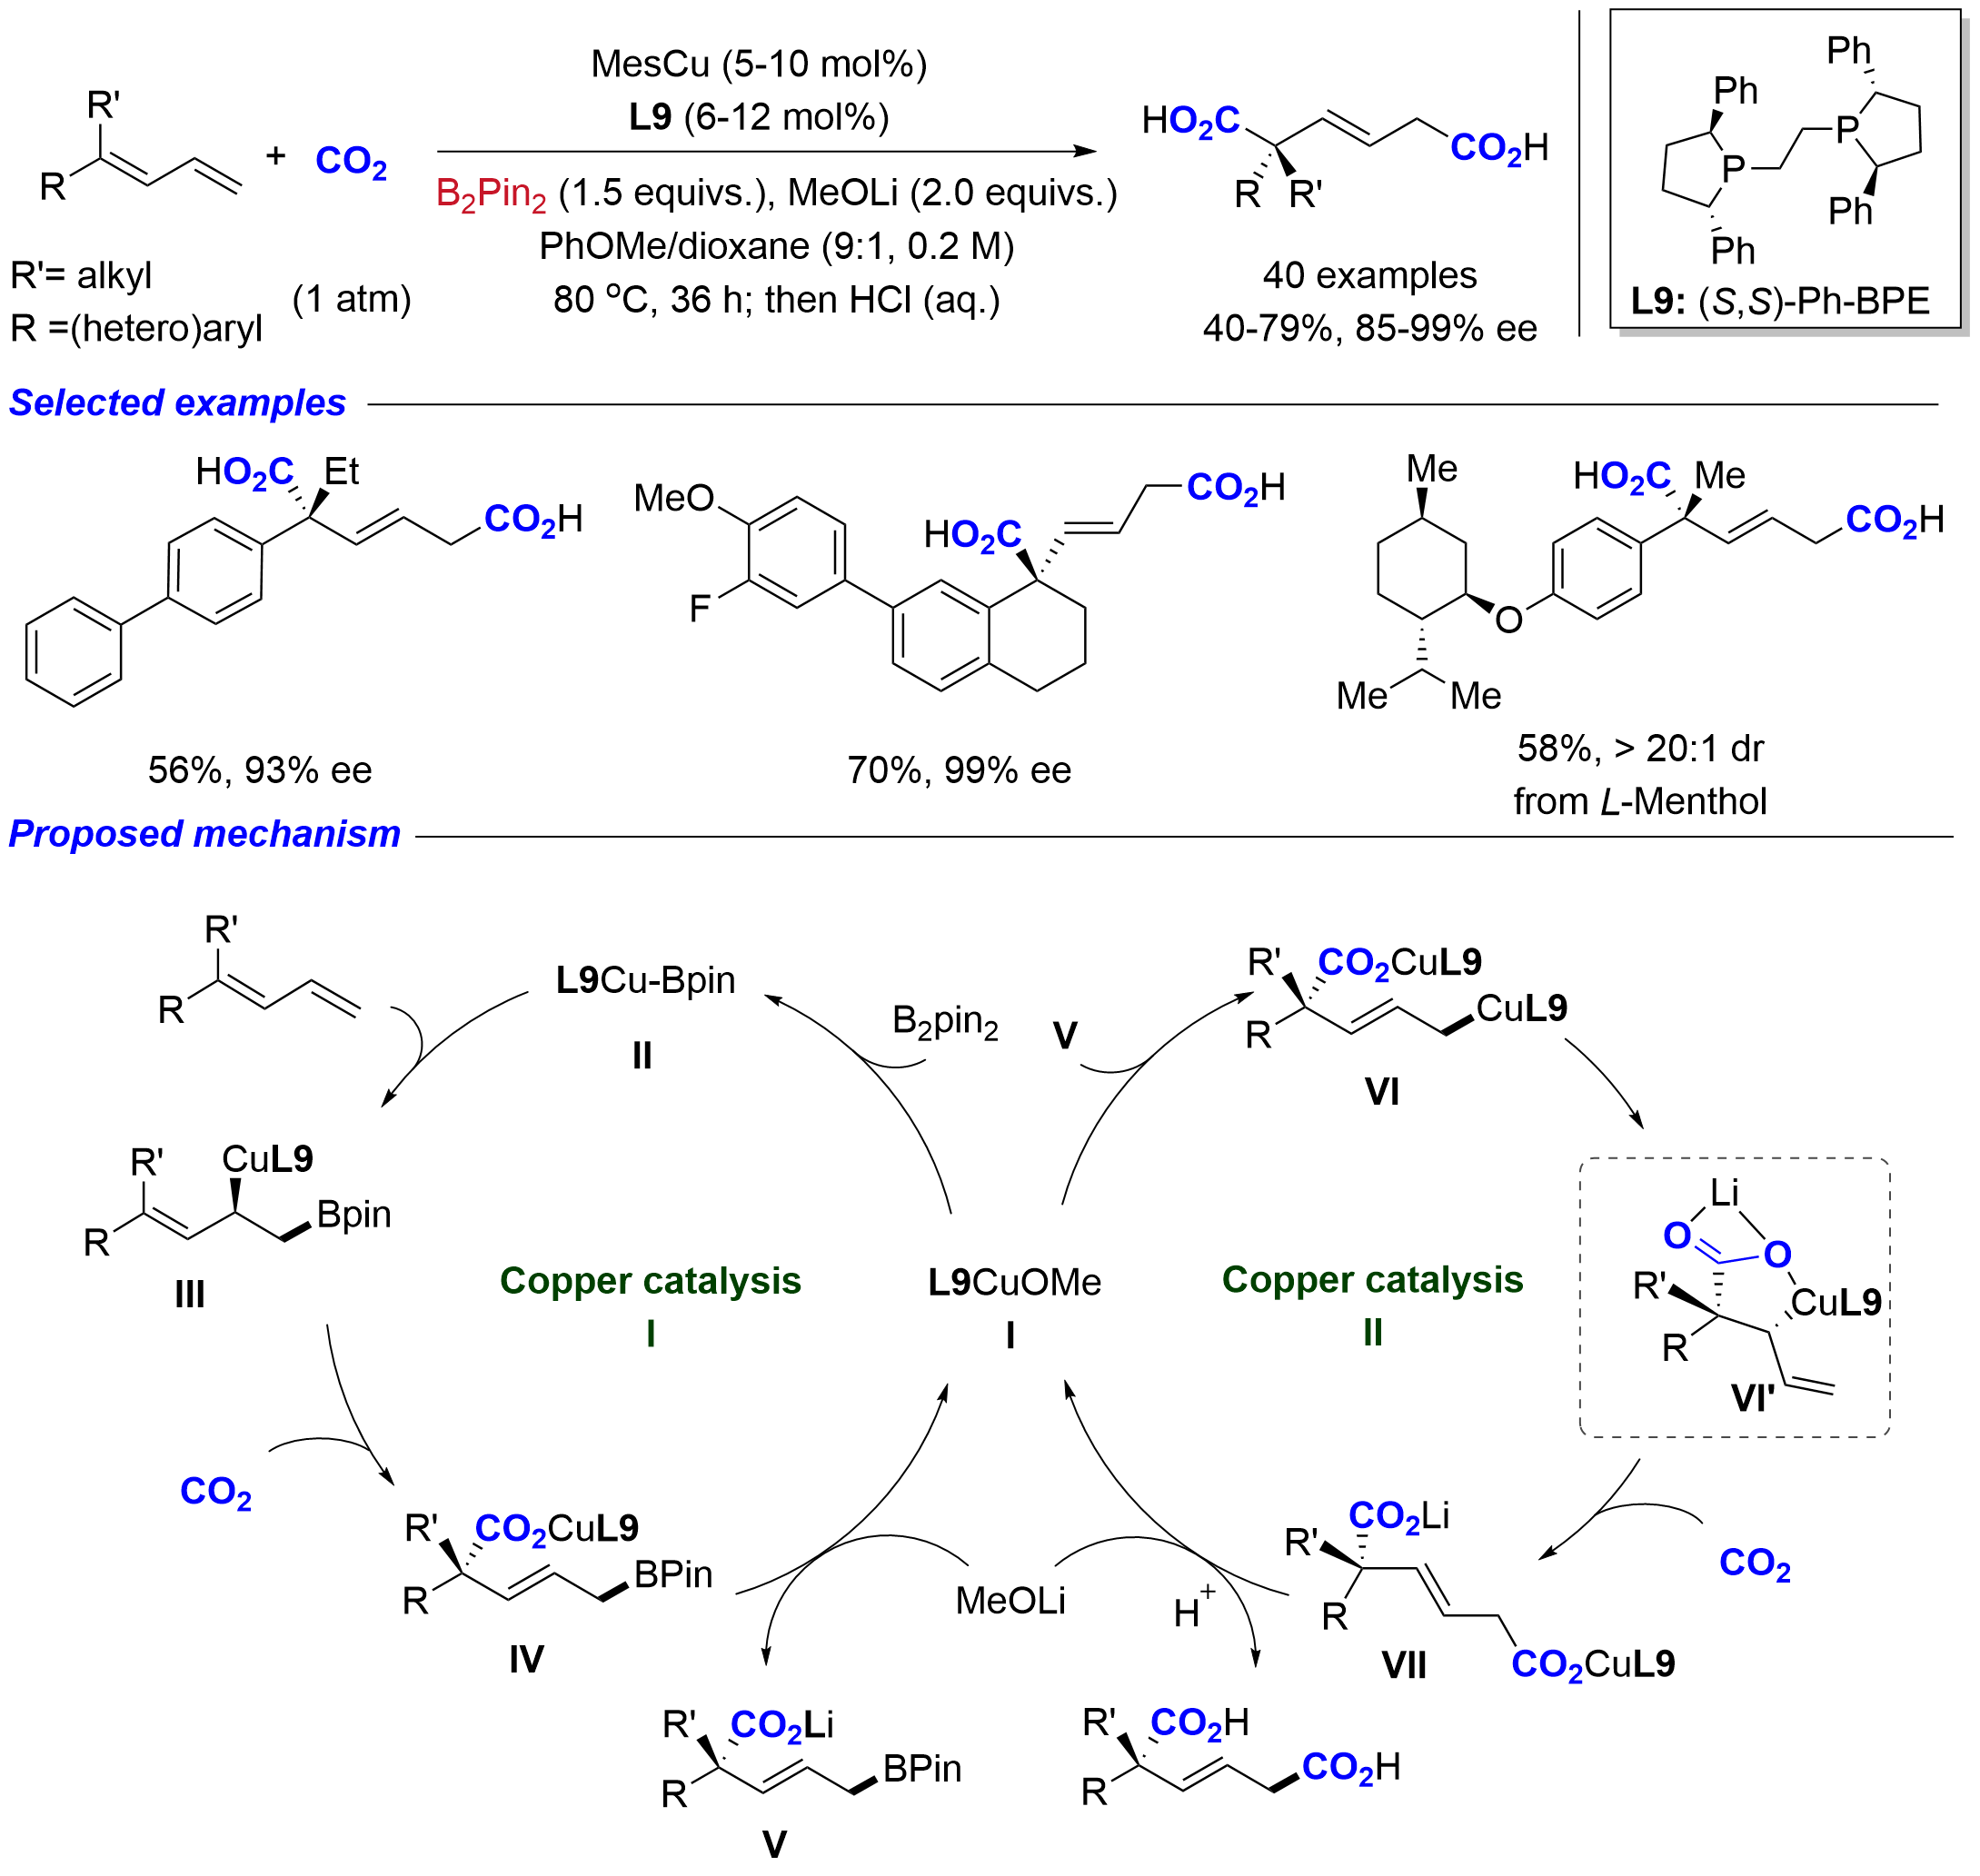
Dicarboxylation of alkenes. In addition to carbo-carboxylation, boracarboxylation and hydrocarboxylation, dicarboxylation provides a fourth distinct strategy for alkene carboxylation, enabling the incorporation of two CO2 molecules into a single alkene substrate. This reaction requires simultaneous control of the regioselectivity of two carboxyl groups and the enantioselectivity of the newly formed stereogenic center, which is particularly challenging for sterically congested quaternary carbon stereocenters. In 2024, Yu, Gui and coworkers reported a copper-catalyzed asymmetric dicarboxylation of 1,3-dienes with CO2. Using MesCu as the catalyst and (S, S)-Ph-BPE as the chiral ligand, dicarboxylic acid derivatives bearing quaternary stereocenters were obtained in up to 79% yield, > 99% ee, > 95:5 regioselectivity, and > 98:2 E/Z selectivity of the resultant double bond (Scheme 11)[27].
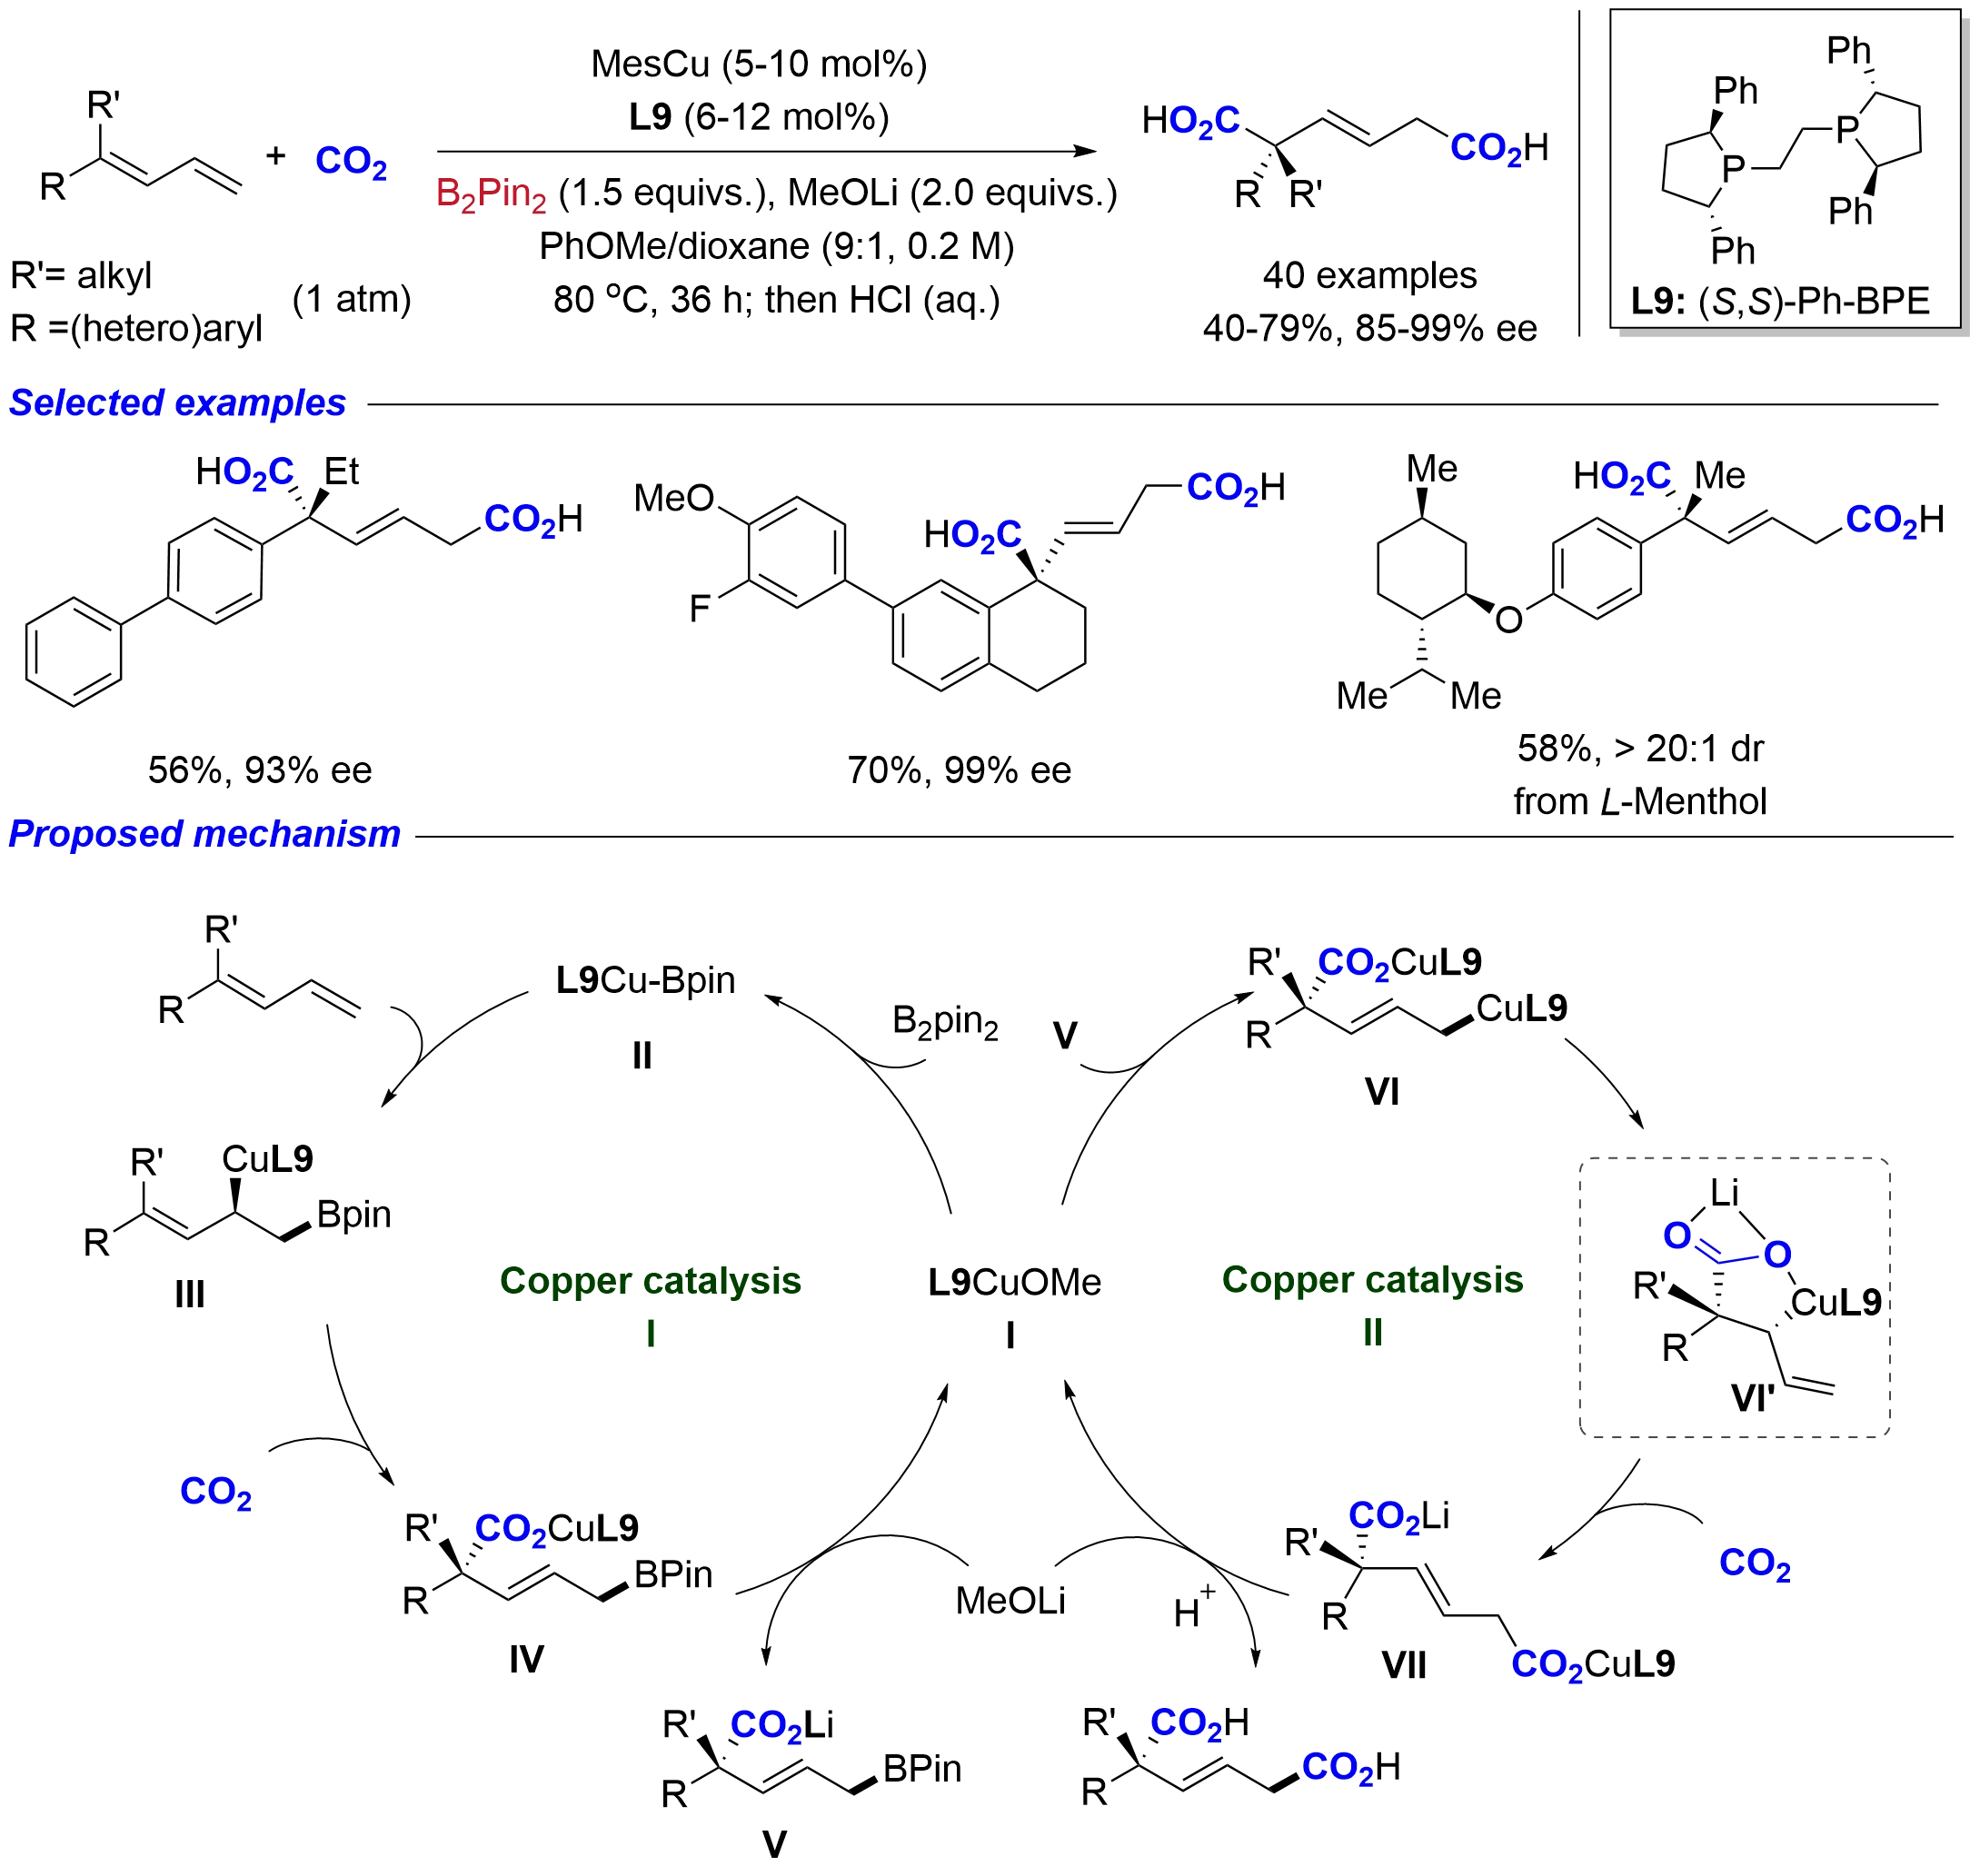
This method features broad substrate scope and good functional group tolerance, and the resulting chiral dicarboxylic acids can be applied to the synthesis of chiral liquid crystalline polyesters and drug-like scaffolds. Mechanistic studies revealed that this reaction proceeds via a copper self-relay catalysis mechanism. The in situ generated Cu(I) complex I reacts with B2pin2 to form the active borylcopper species II, which undergoes 1,2-insertion into the terminal double bond of 1,3-diene to generate boryl-substituted allylcopper intermediate III. Carboxylation of III with CO2 at the benzylic position affords intermediate IV, which undergoes ligand exchange with base to regenerate Cu(I) complex I and release carboxylated allyl boronate ester V. Transmetalation between V and Cu(I) complex I generates allylcopper intermediate VI, which exists in equilibrium with its isomer VI’. Given that the carboxylate group could coordinate with copper to form a more favorable five-membered metallacyclic intermediate VI’, this species selectively reacts with CO2 at the terminal position to furnish dicarboxylate VII, which affords the target product upon acidic workup. In this process, the carboxylate group introduced in the first step serves as a directing group, effectively controlling the regioselectivity and E/Z-selectivity of the second carboxylation.
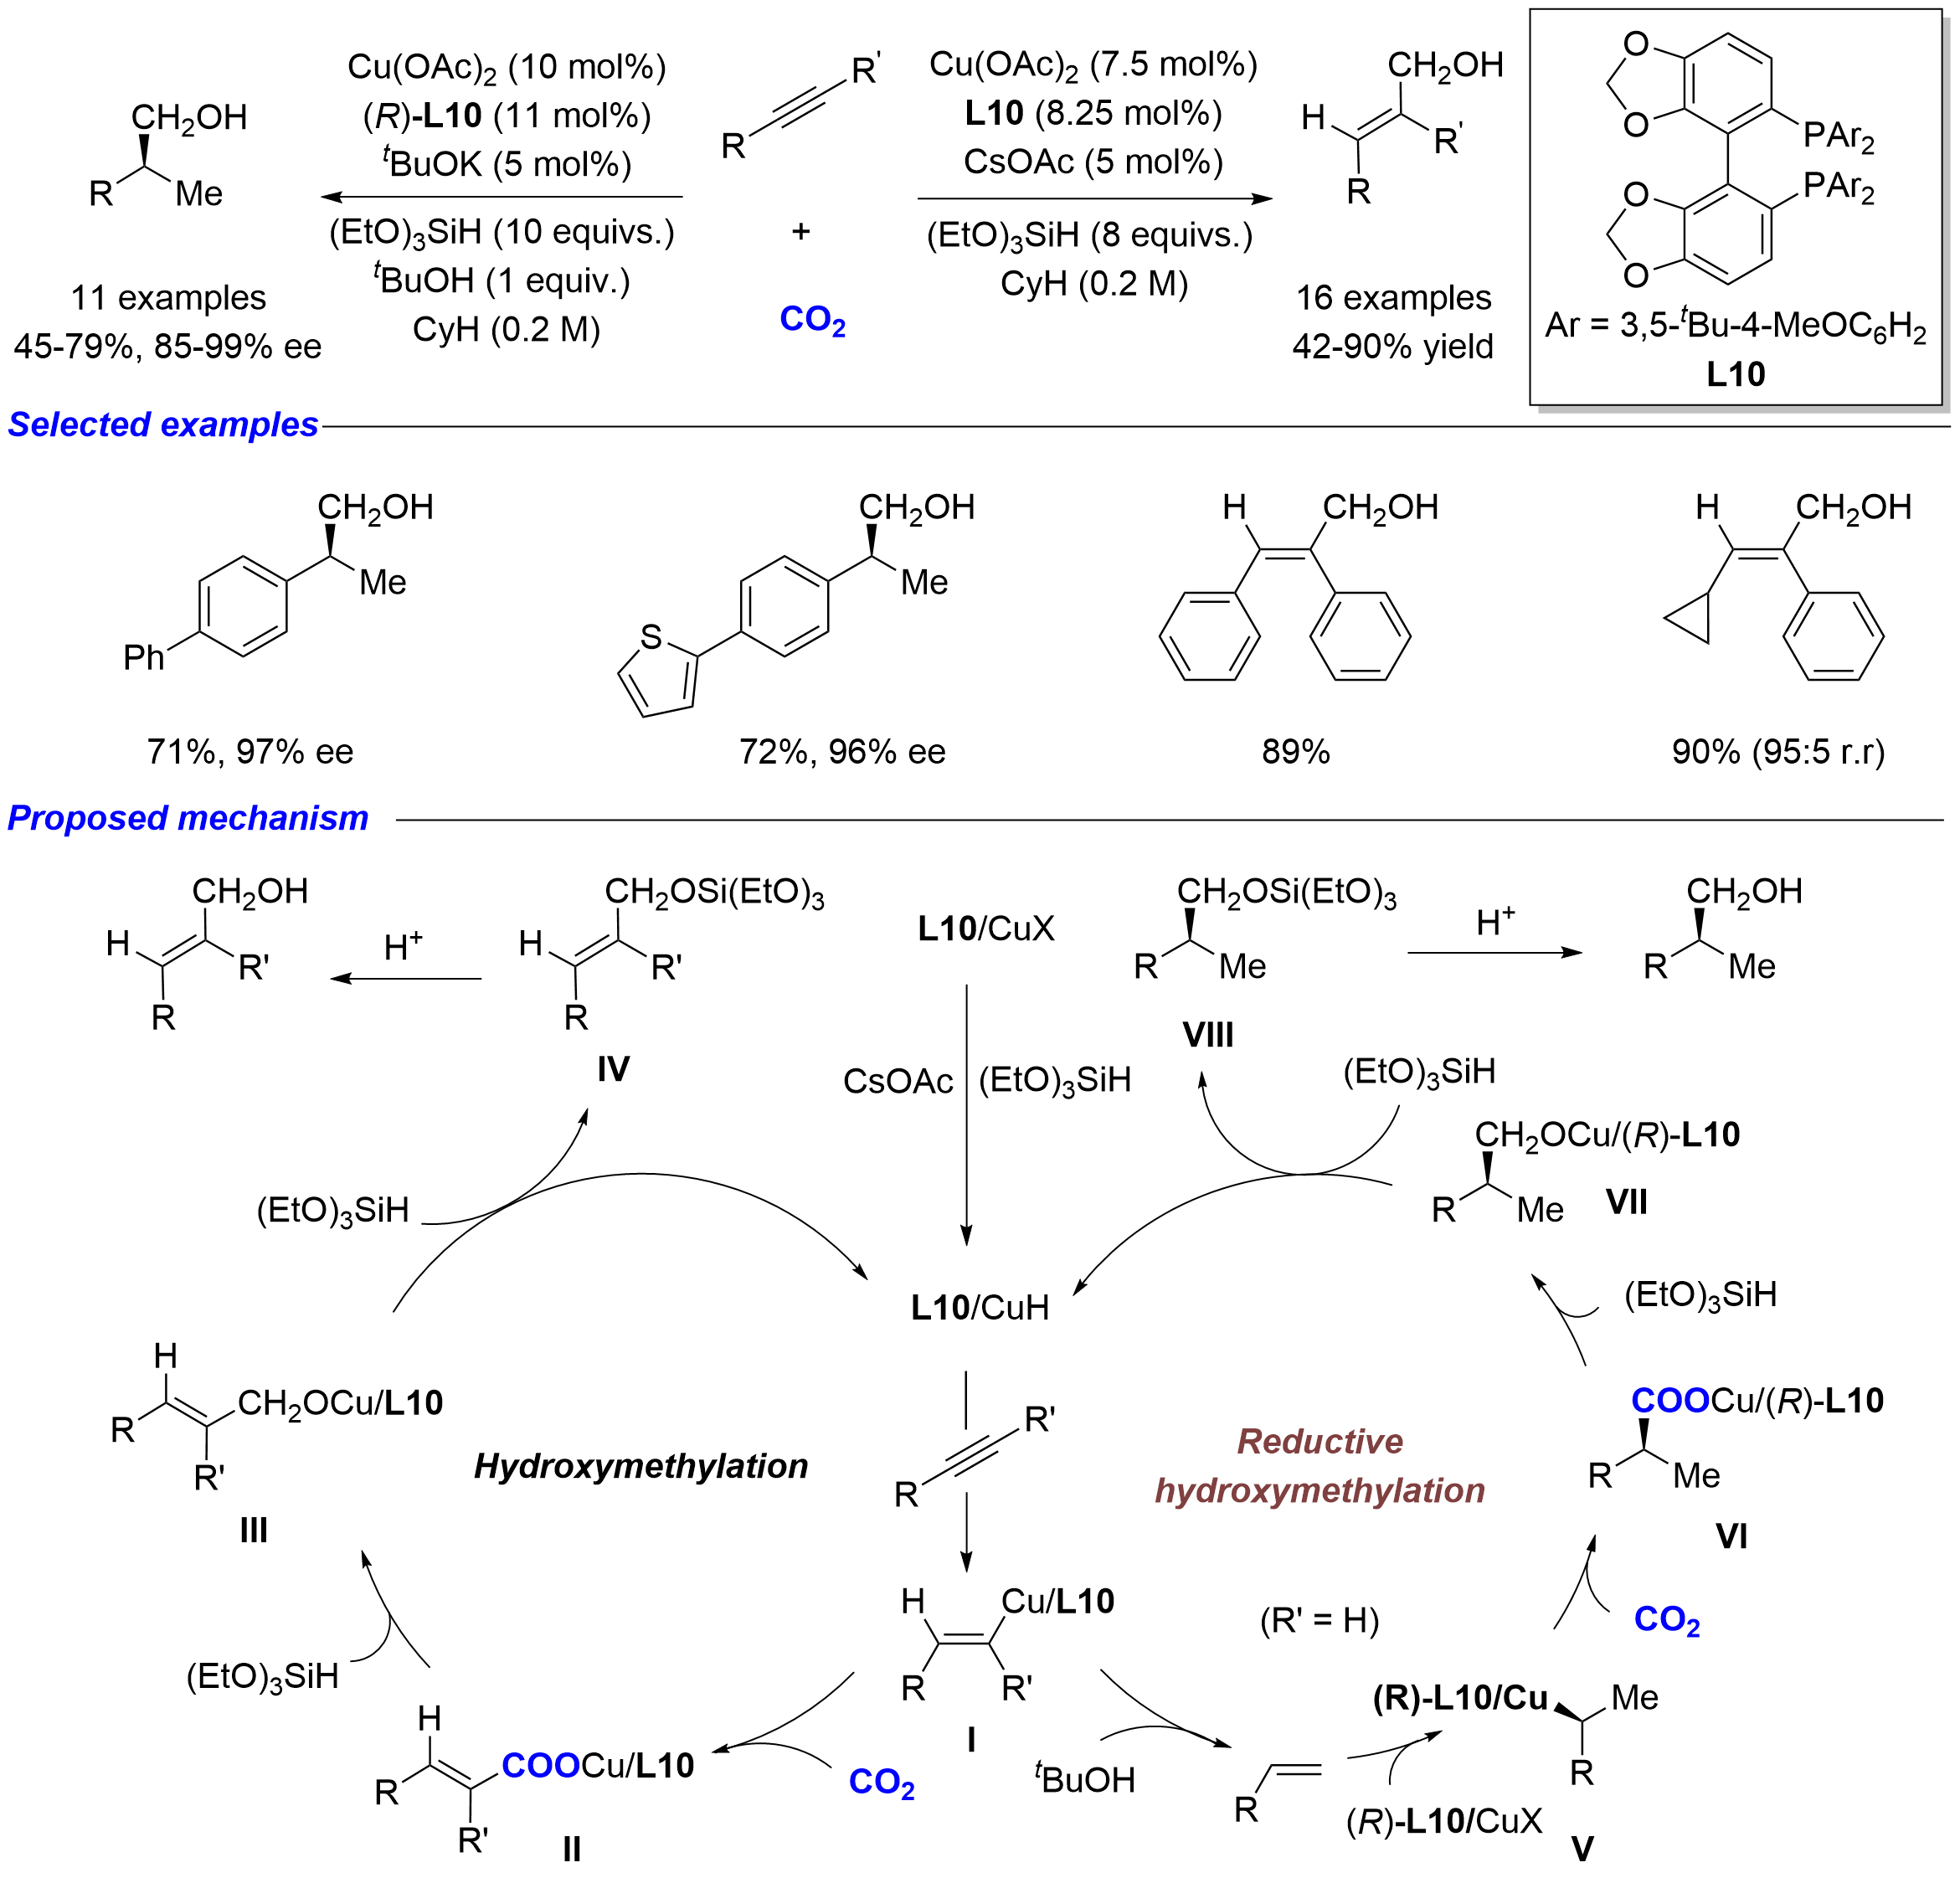
CO2-involved hydroxymethylation of alkynes. The linear structure and tunable π-electron system of alkynes provide a unique platform for asymmetric carboxylation strategies. In addition to the direct construction of chiral carboxylic acids, alkynes can also serve as versatile substrates for the synthesis of chiral alcohols through CO2-involved hydroxymethylation, wherein CO2 functions as a C1 synthon and is ultimately reduced to a hydroxymethyl group. In 2021, Ma, He and coworkers developed a copper-catalyzed selective hydroxymethylation of alkynes with CO2 (Scheme 12)[28].
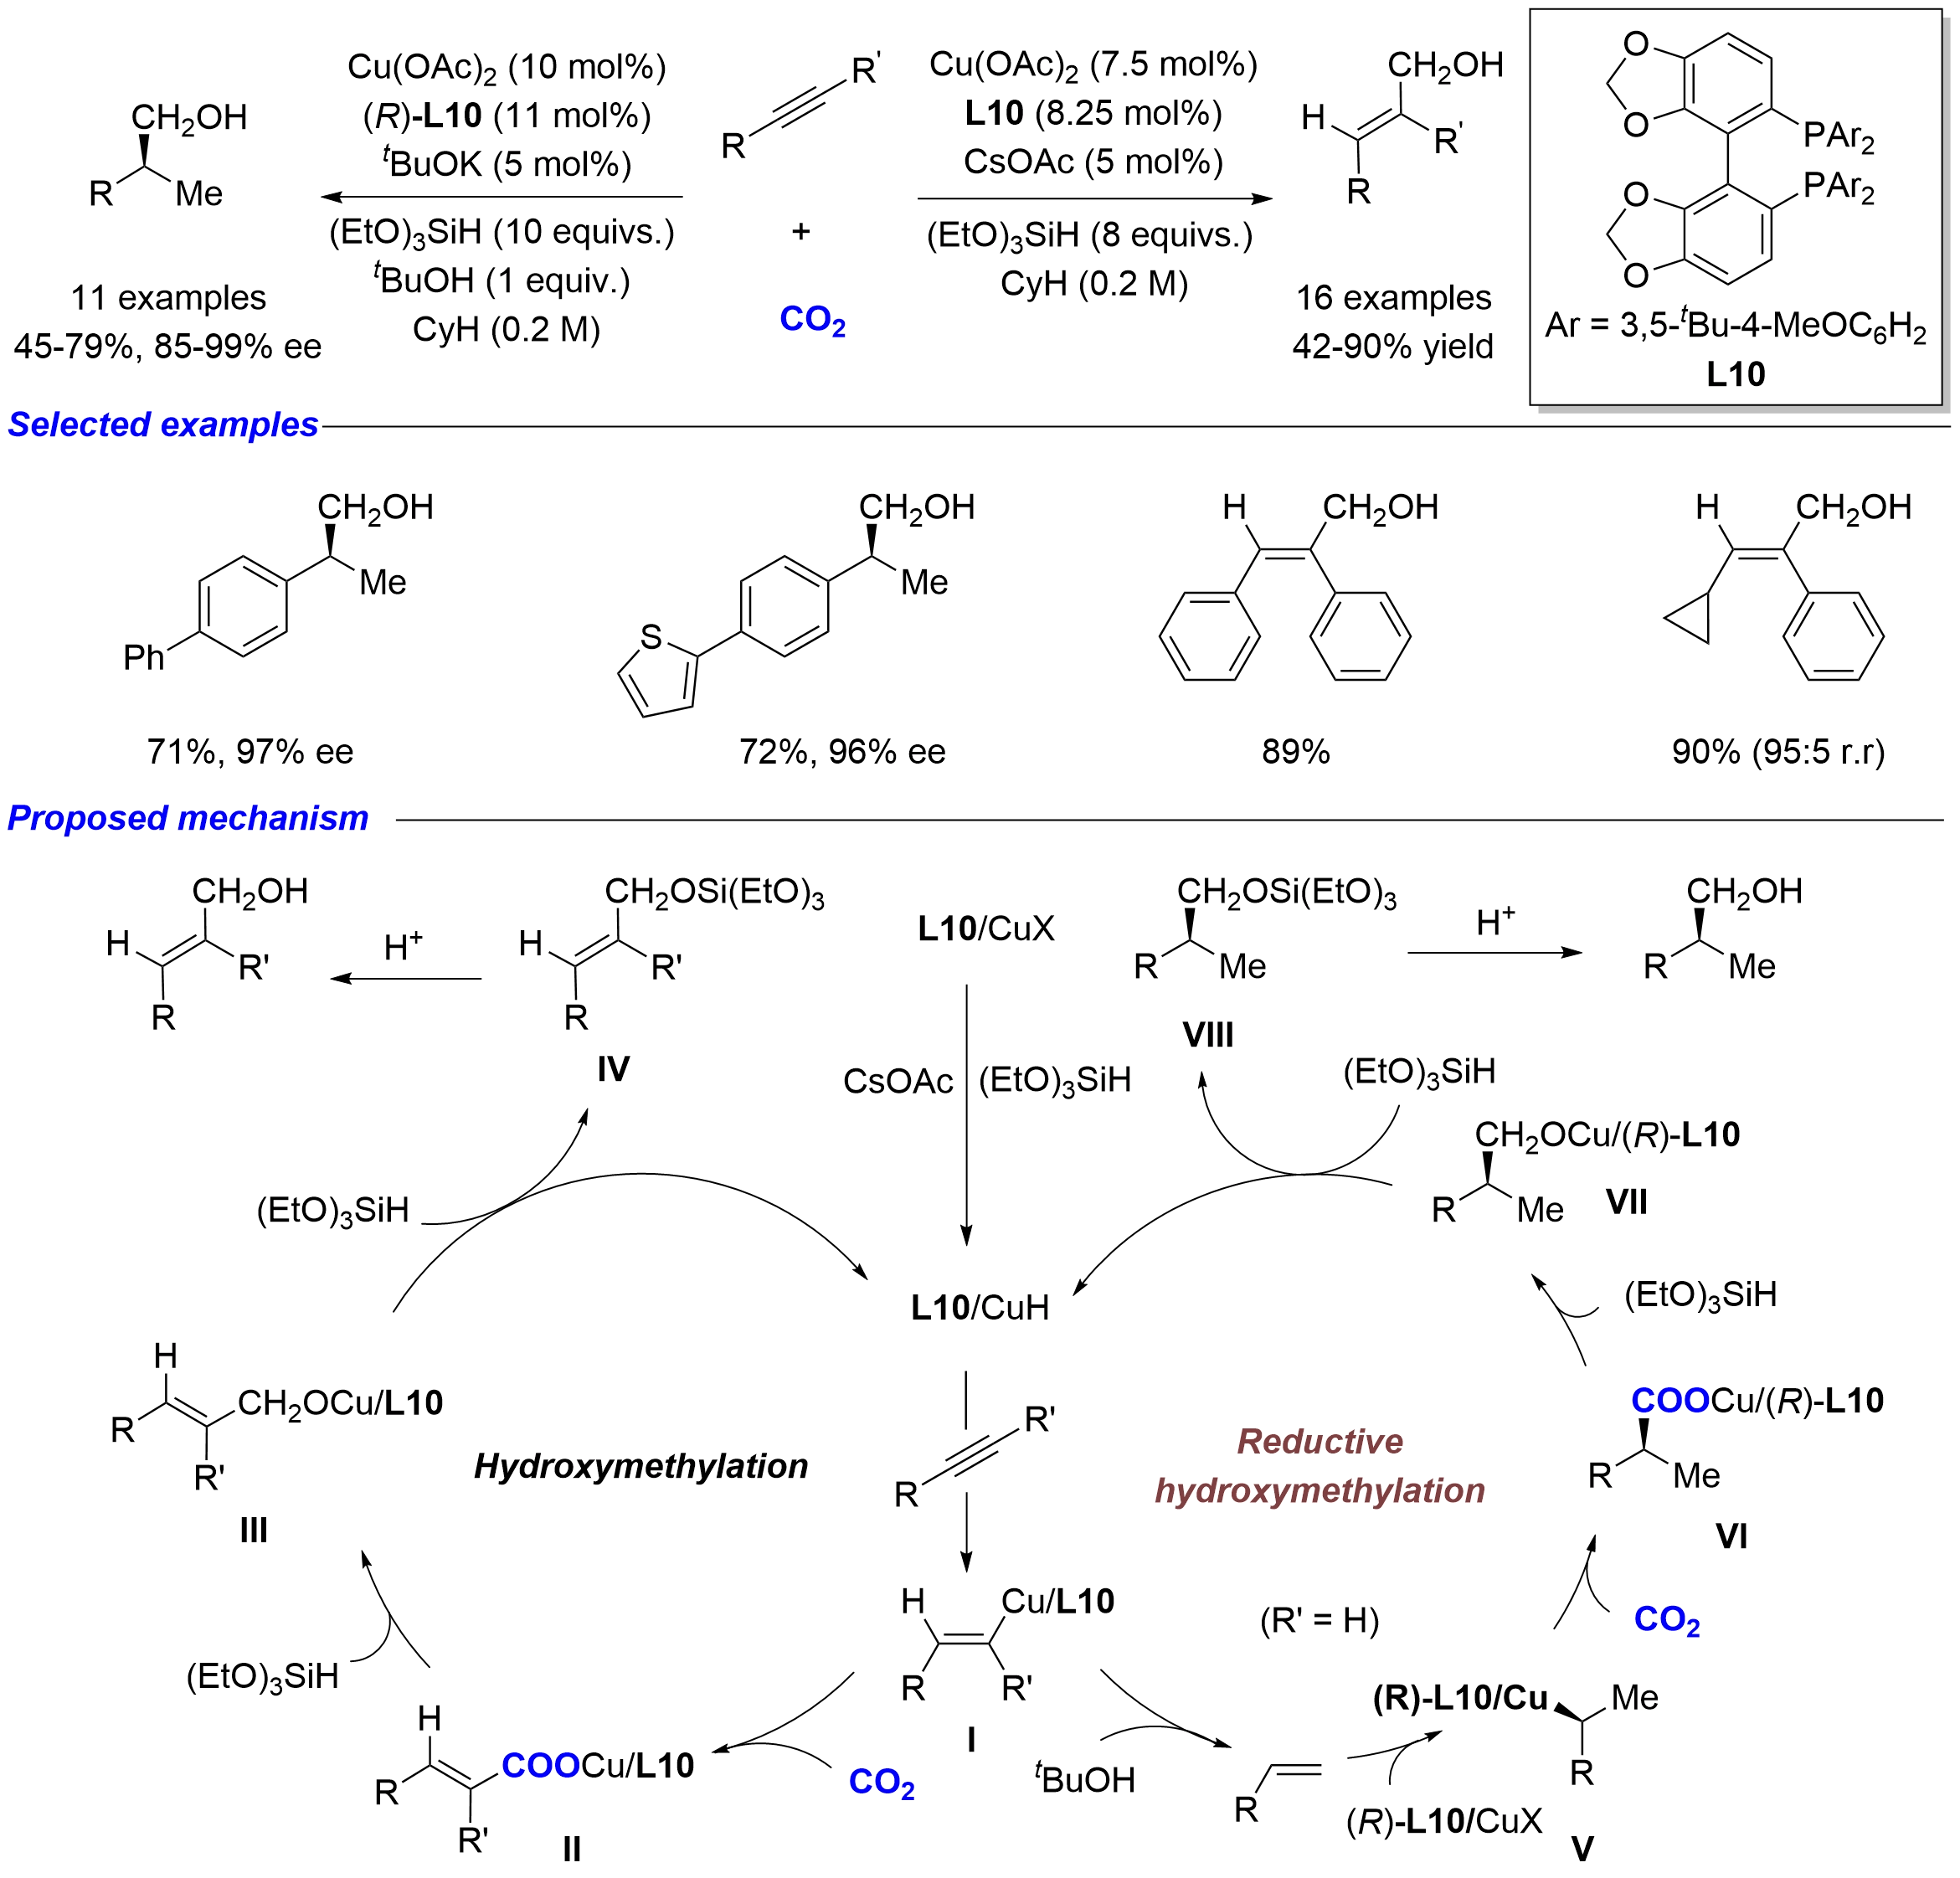
Using Cu(OAc)2 as the catalyst, DTBM-SEGPHOS (L10) as the chiral ligand, triethoxysilane as the hydride source, and 1 atm CO2 in cyclohexane, two distinct transformations, namely direct hydroxymethylation and reductive hydroxymethylation, could be achieved by controlling the presence or absence of a proton source. This strategy afforded allylic alcohols or chiral benzylic alcohols with high regio-, stereo-, and enantioselectivity. Mechanistic studies revealed that both transformations originate from the hydrocupration of the alkyne by a copper hydride species, generating a common vinylcopper intermediate I. In the absence of the proton source, intermediate I is directly captured by CO2 to form copper carboxylate species II, which undergoes reduction to afford intermediate III. Subsequent transmetalation with a hydrosilane regenerates the CuH species and ultimately delivers the allylic alcohol product. On the other hand, in the presence of the proton source tBuOH, the vinylcopper intermediate I undergoes preferential protonation to generate an alkene, which then undergoes a second hydrocupration to form the benzylic copper species V. Intermediate V subsequently undergoes carboxylation with CO2 to afford copper carboxylate species VI, which upon further reduction yields the homobenzylic alcohol product. The presence or absence of the protic additive dictates whether the vinylcopper species undergoes protonation or hydroxymethylation, ultimately determining the product.
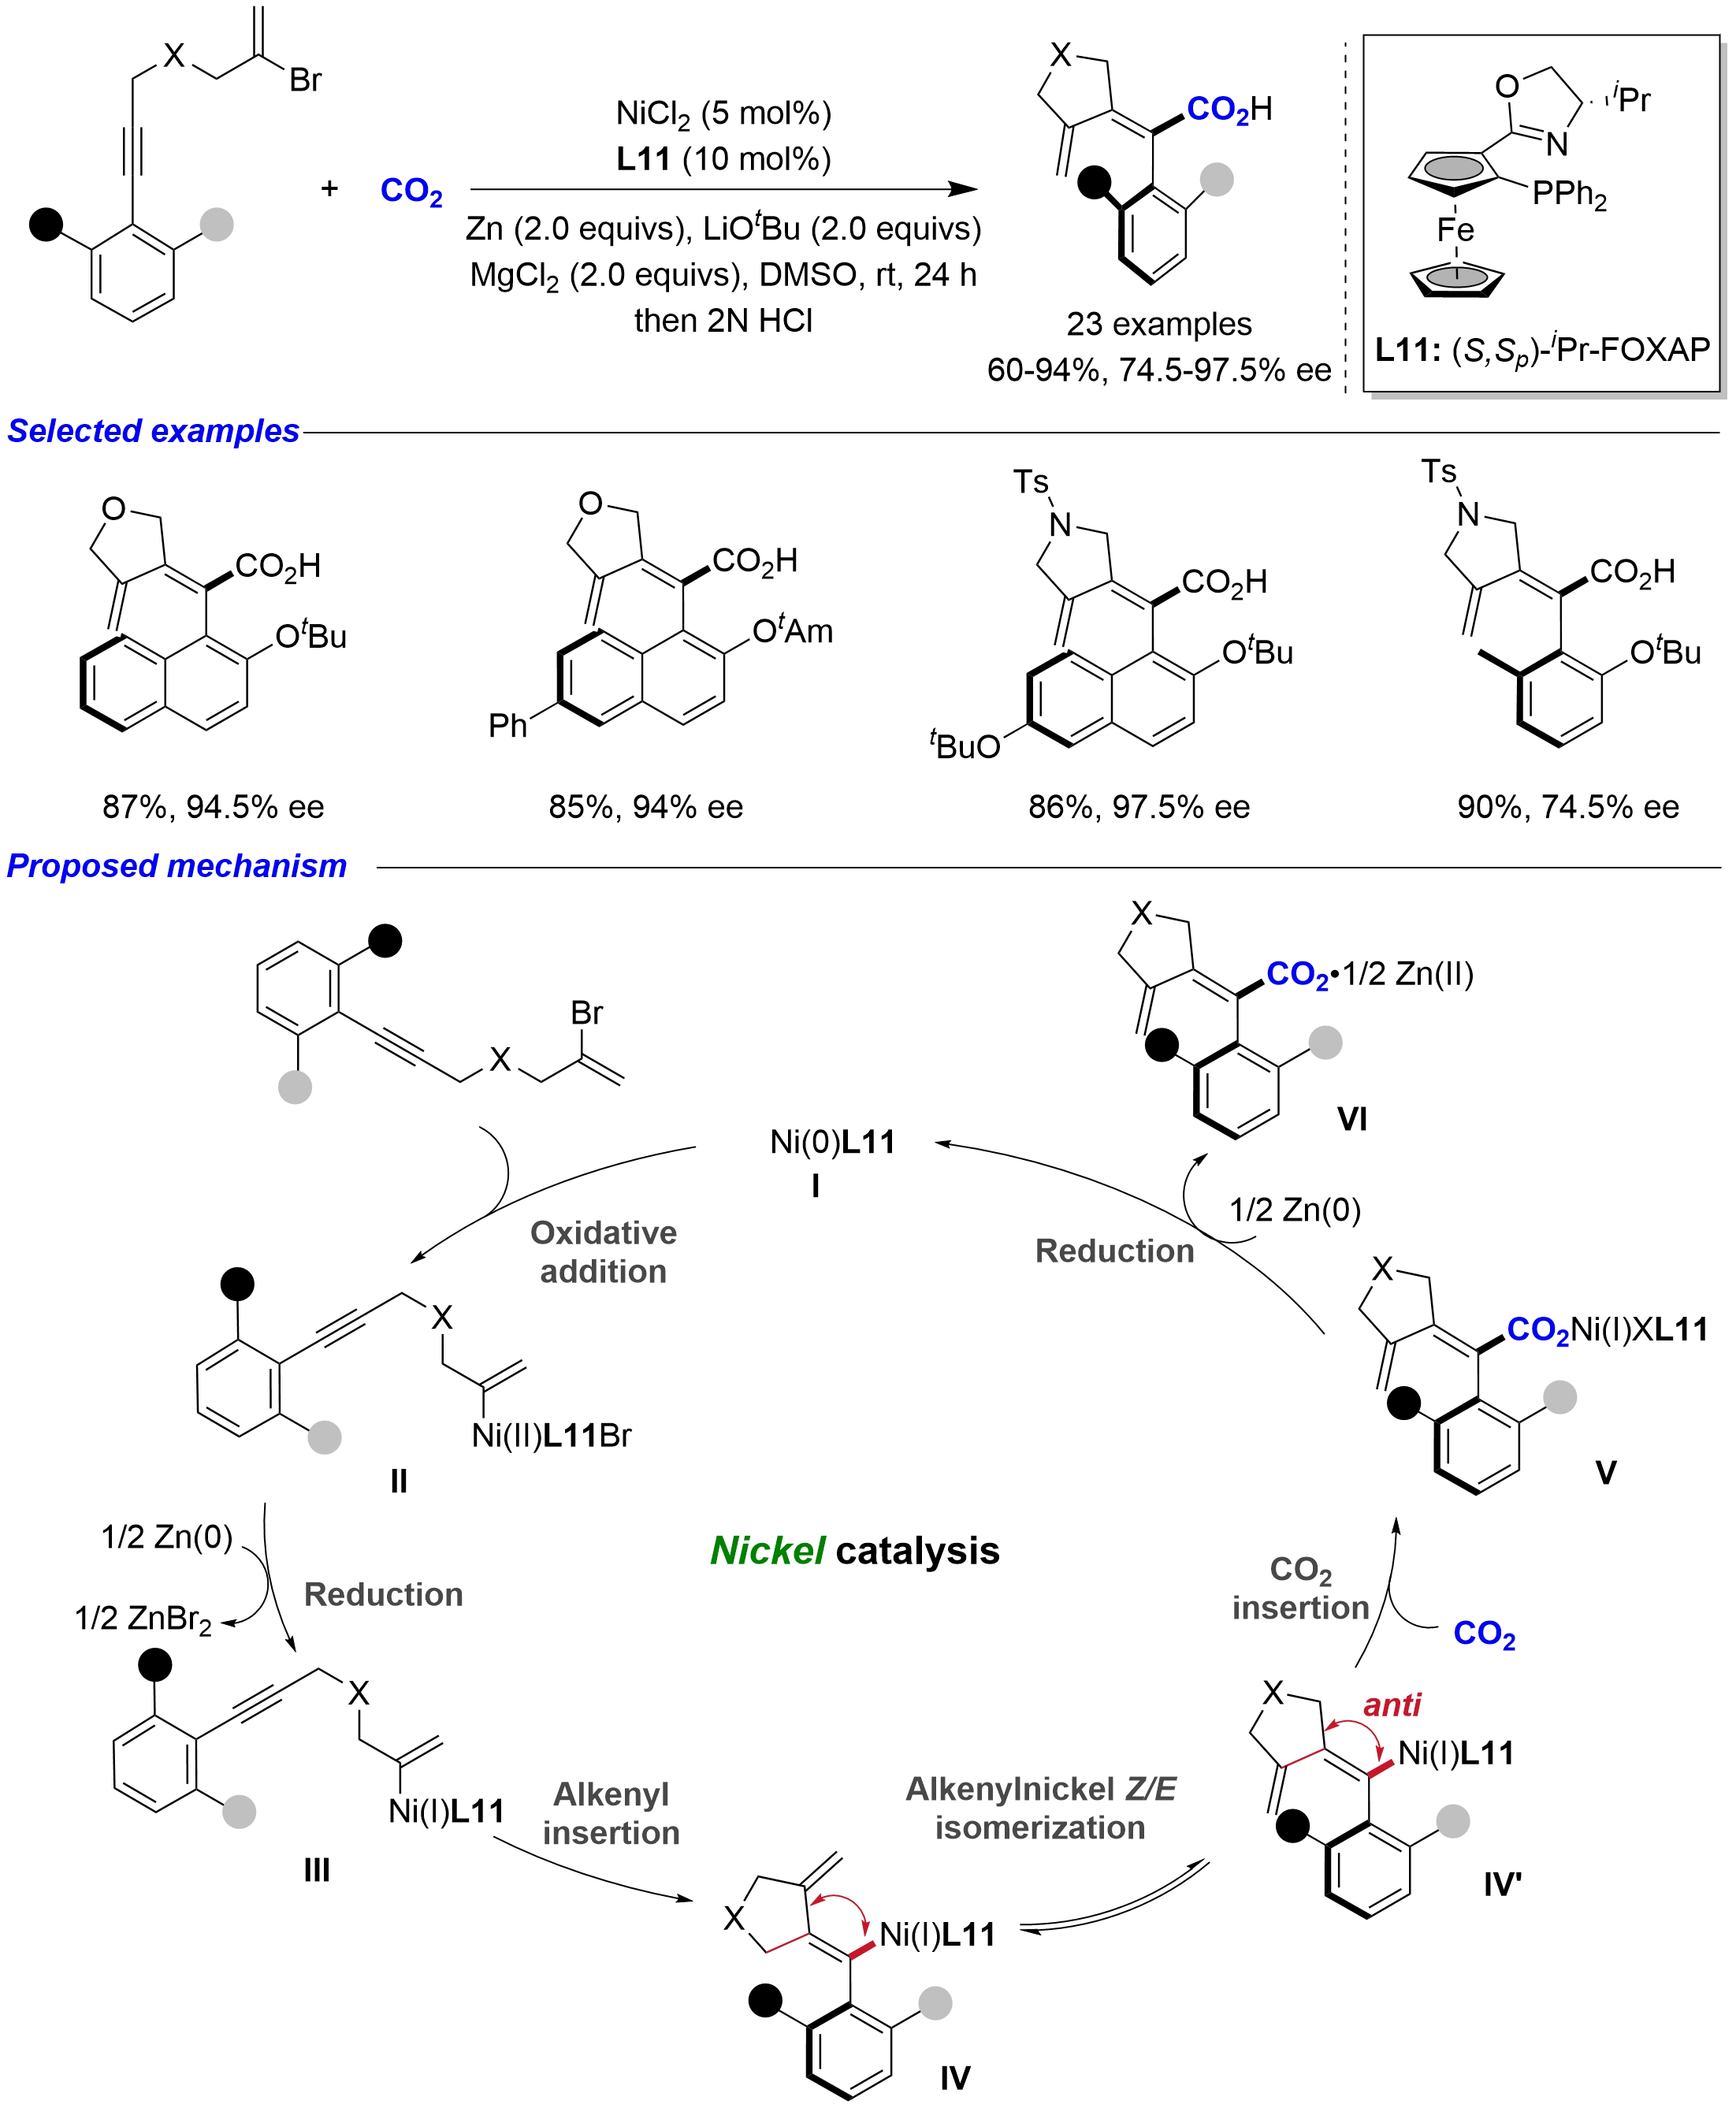
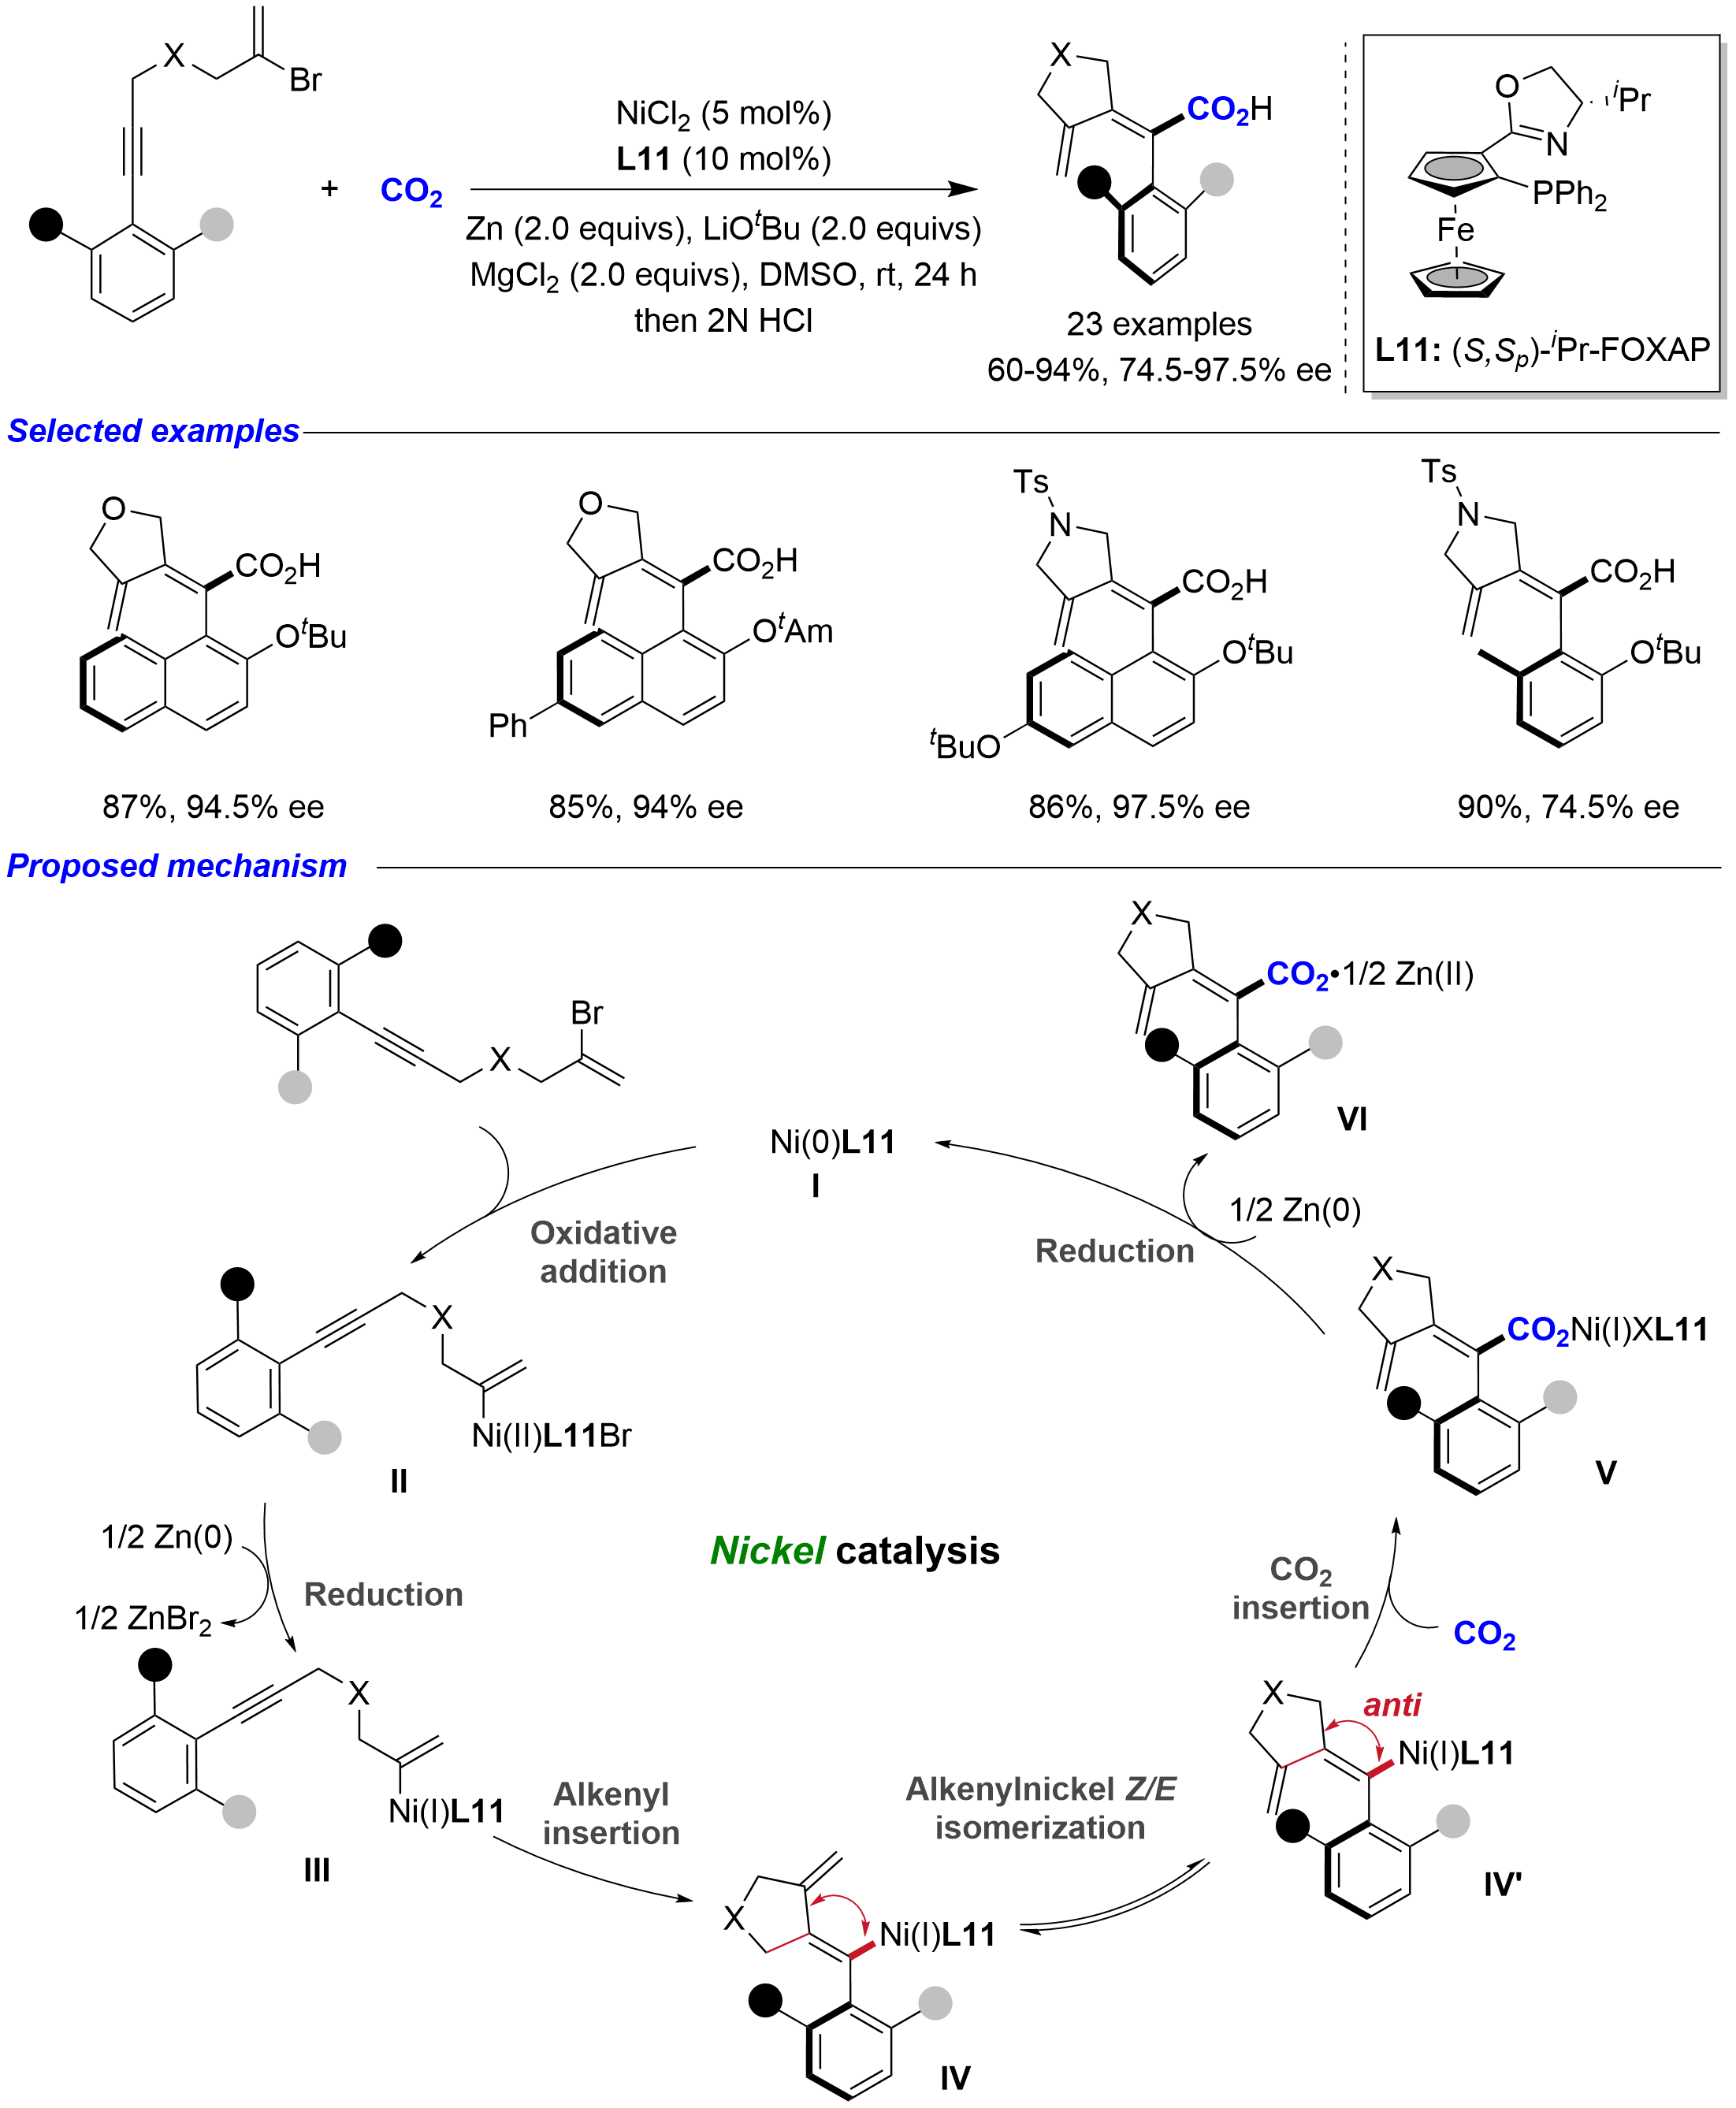
Carbo-carboxylation of alkynes. In 2025, Yu, Zhou, Gui and coworkers reported an elegant chiral nickel-catalyzed atroposelective carbo-carboxylation of alkynes with CO2 (Scheme 13)[29]. Using NiCl2 and a chiral ferrocenyl phosphine-oxazoline (FOXAP) ligand, an intramolecular cyclization-carboxylation cascade efficiently constructed axially chiral styrene-type carboxylic acids bearing tetrahydrofuran or pyrrolidine frameworks with high chemo-, regio-, stereo-, and enantioselectivity. The resulting axially chiral carboxylic acids could serve as chiral ligands for chiral Co-catalyzed C-H activation of indole compounds. Mechanistic studies including DFT calculations revealed that the active Ni(0) species I first undergoes oxidative addition with the alkynyl bromide substrate to generate the vinyl Ni(II)L11 intermediate II. Subsequently, II is reduced by zinc powder to afford the vinyl Ni(I)L11 species III. Intermediate III undergoes enantioselective 2,1-syn insertive cyclization with the intramolecularly tethered alkyne to generate the chiral vinyl Ni(I)L11 intermediate IV. This intermediate then undergoes E/Z isomerization to afford the thermodynamically more stable Z-configured vinyl Ni(I)L11 intermediate IV′. Subsequently, IV′ undergoes nucleophilic addition to CO2 to generate the nickel carboxylate species V. Further reduction of V by zinc powder releases the zinc carboxylate VI and regenerates the Ni(0) species I, with VI affording the target axially chiral carboxylic acid product upon protonation during workup.
2.2 Asymmetric carboxylation of organic electrophiles with CO2
Organic (pseudo)halides and related electrophilic substrates constitute a broad class of widely available and structurally diverse coupling partners. Unlike alkenes and alkynes, which react with CO2 through unsaturated bonds, these substrates can undergo oxidative addition with a transition metal to form a metal-carbon bond, thereby enabling enantioselective carbon-carbon bond formation and providing an efficient alternative route to chiral carboxylic acids and their derivatives. In recent years, significant breakthroughs have been achieved in asymmetric carboxylation reactions based on a wide range of organic electrophiles, including aryl and alkenyl halides, benzyl ammonium salts, diaryliodonium salts, propargylic carbonates, and heterobiaryl derivatives.
In 2022, Li and coworkers designed a novel chiral 2,2′-bipyridine ligand, Me-SBpy (L12), for the chiral nickel-catalyzed enantioconvergent carboxylation of racemic benzylic (pseudo)halides (Scheme 14)[30]. Using NiCl2(PCy3)2 as the catalyst precursor, Mn powder as the reductant, and 1 atm CO2, the reaction proceeded in DMF/2-MeTHF/MeOH at -10 °C for 16 h, converting racemic benzylic (pseudo)halides, such as ammonium salts, bromides, or chlorides, into α-chiral arylpropionic acids in up to 95% yield with up to 98% ee. The synthesis of the profen family of nonsteroidal anti-inflammatory drugs, such as (S)-naproxen, (S)-ketoprofen, and (S)-fenoprofen was successfully realized. Mechanistic studies revealed that manganese powder first reduces the Ni(II) precursor and undergoes ligand exchange with the chiral SBpy ligand to generate the catalytically active chiral Ni0 species I. Subsequently, I undergoes oxidative addition with the benzyl (pseudo)halides to form the SBpy-coordinated benzyl Ni(II) intermediate II. Intermediate II can establish an equilibrium with dissociated Ni(I)X and a benzyl radical, which is crucial for substrate racemization and enantioconvergence. Intermediate II is then reduced by manganese powder to the Ni(I) species III. Insertion of CO2 into III generates the Ni(I) carboxylate IV, a step that can be facilitated by in situ generated manganese salts such as MnCl2 acting as Lewis acids. Finally, IV is further reduced by manganese powder to regenerate the Ni0 species I and release the manganese carboxylate V, which upon acidification affords the target chiral carboxylic acid. Notably, Yu and co-workers provided strong theoretical support for the C-Ni bond homolysis/recombination enantioconvergent mechanism through DFT calculations[31].
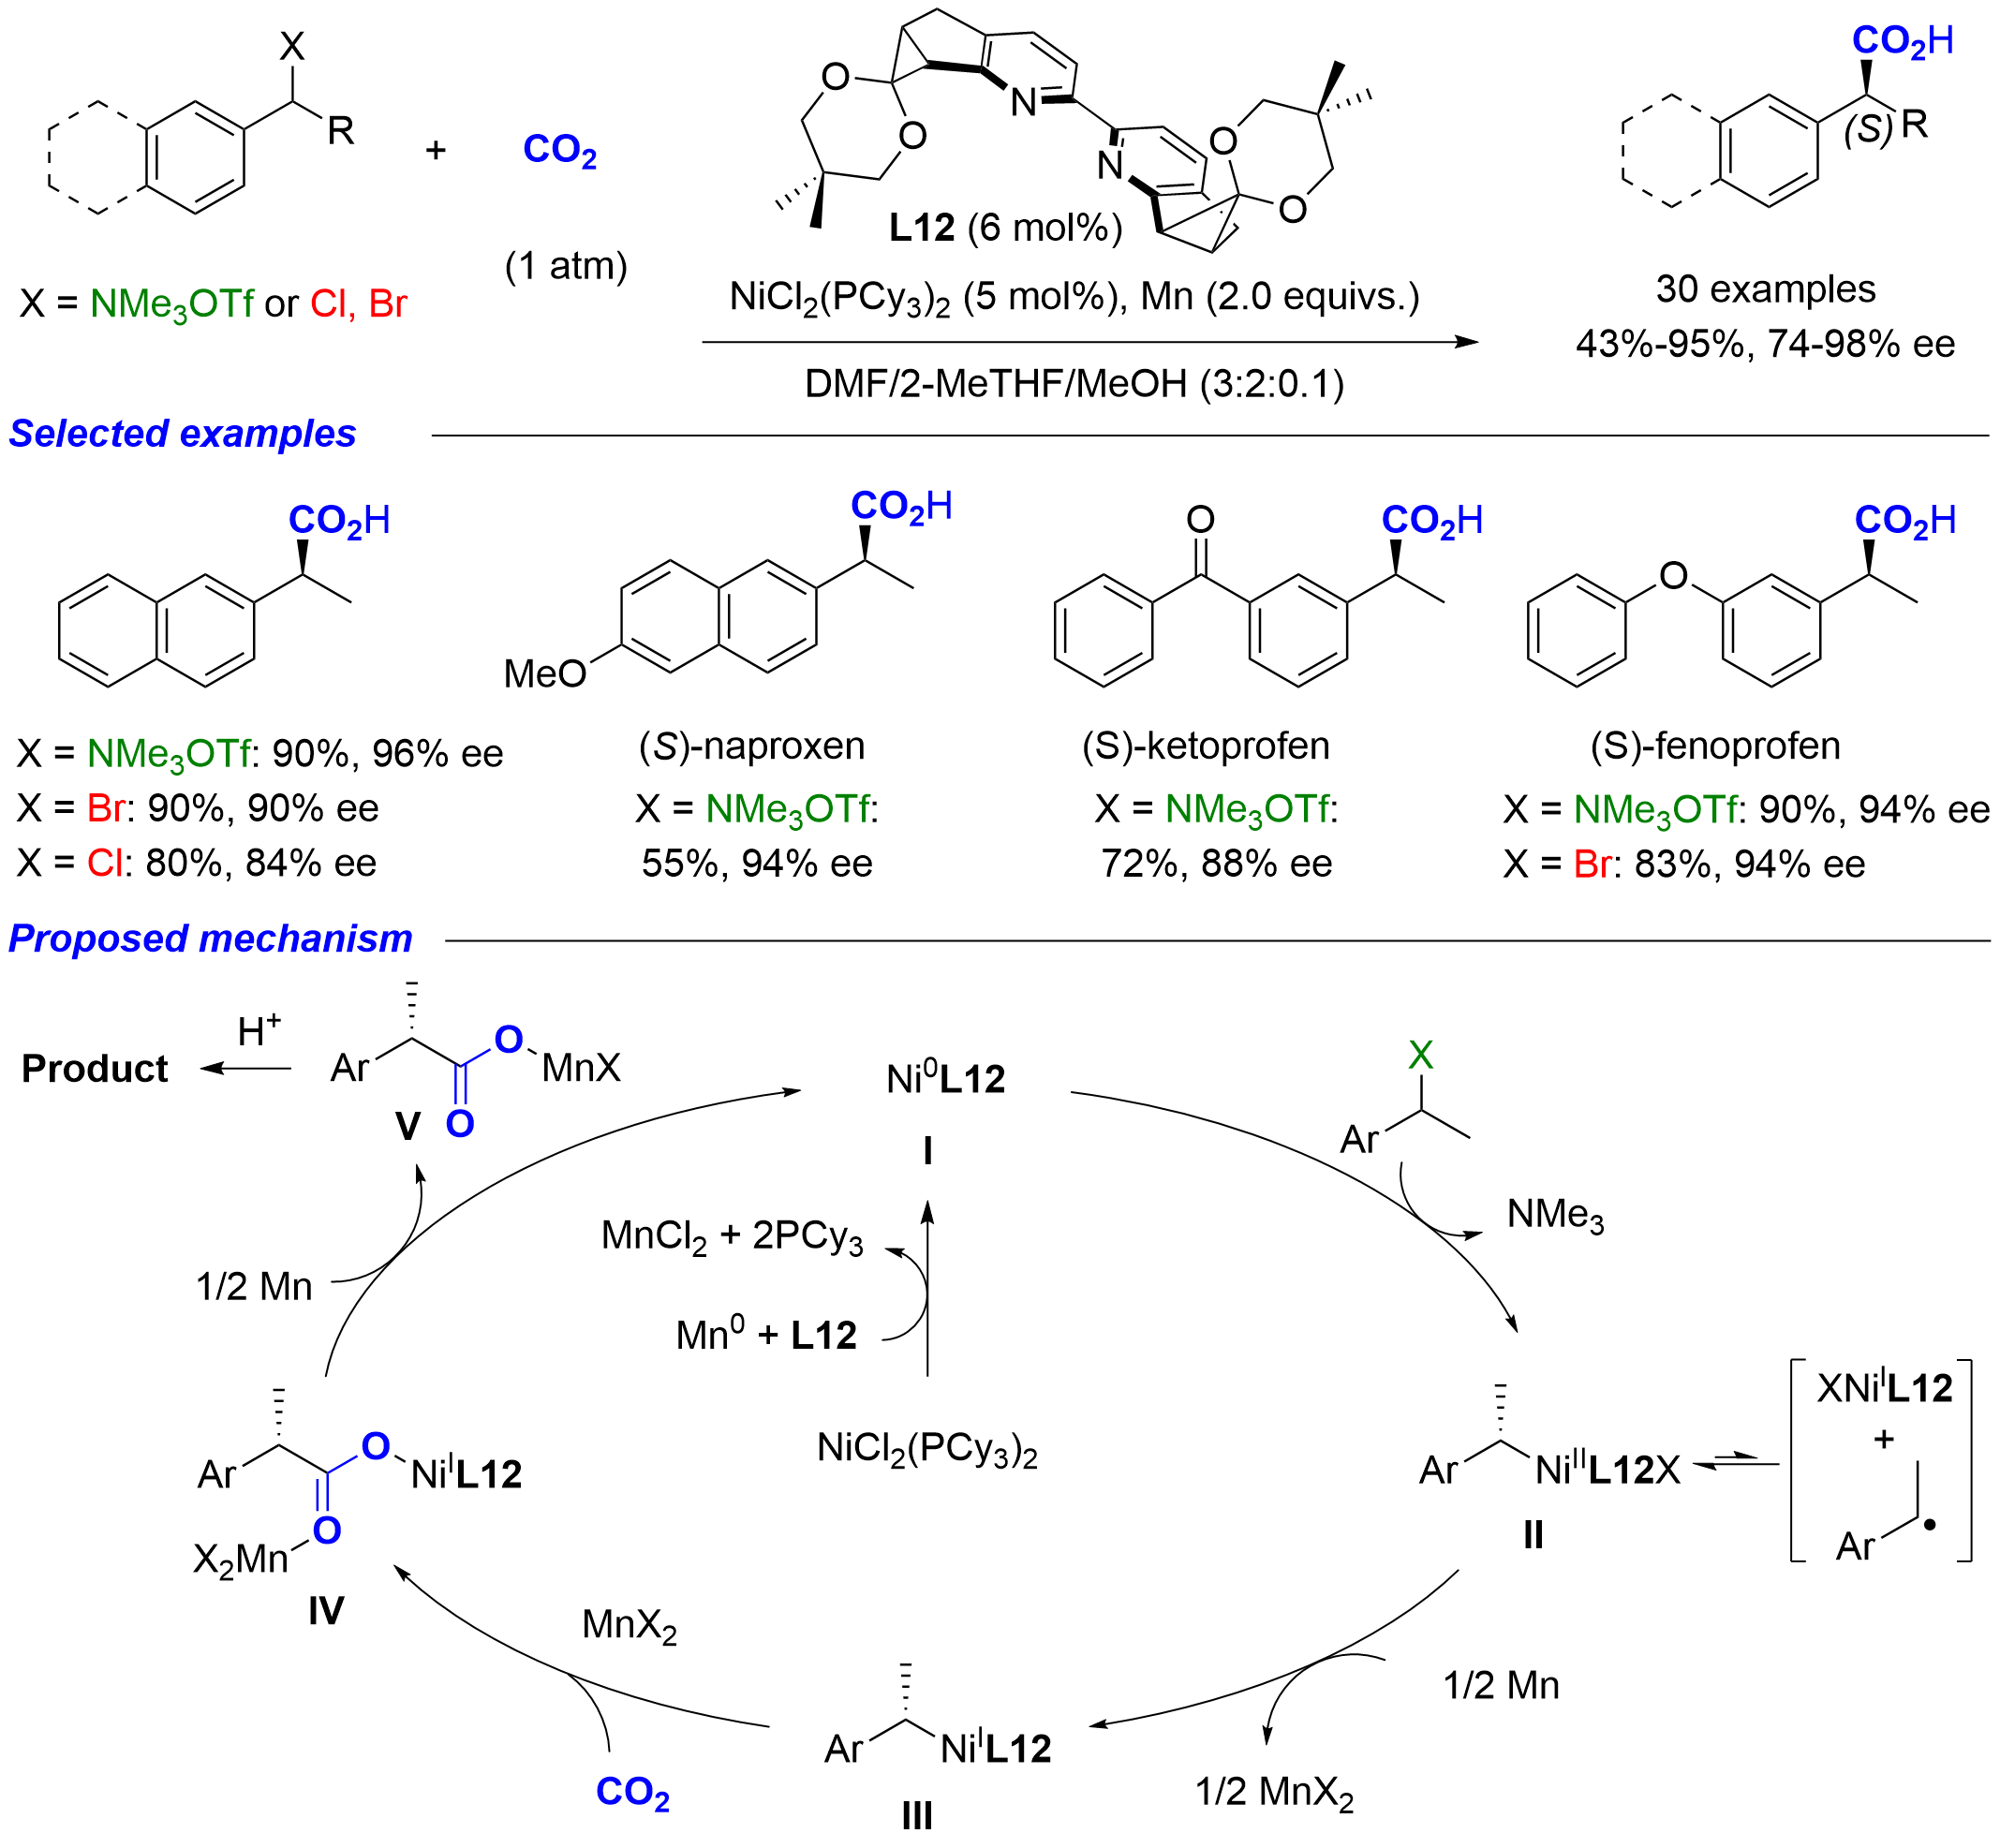
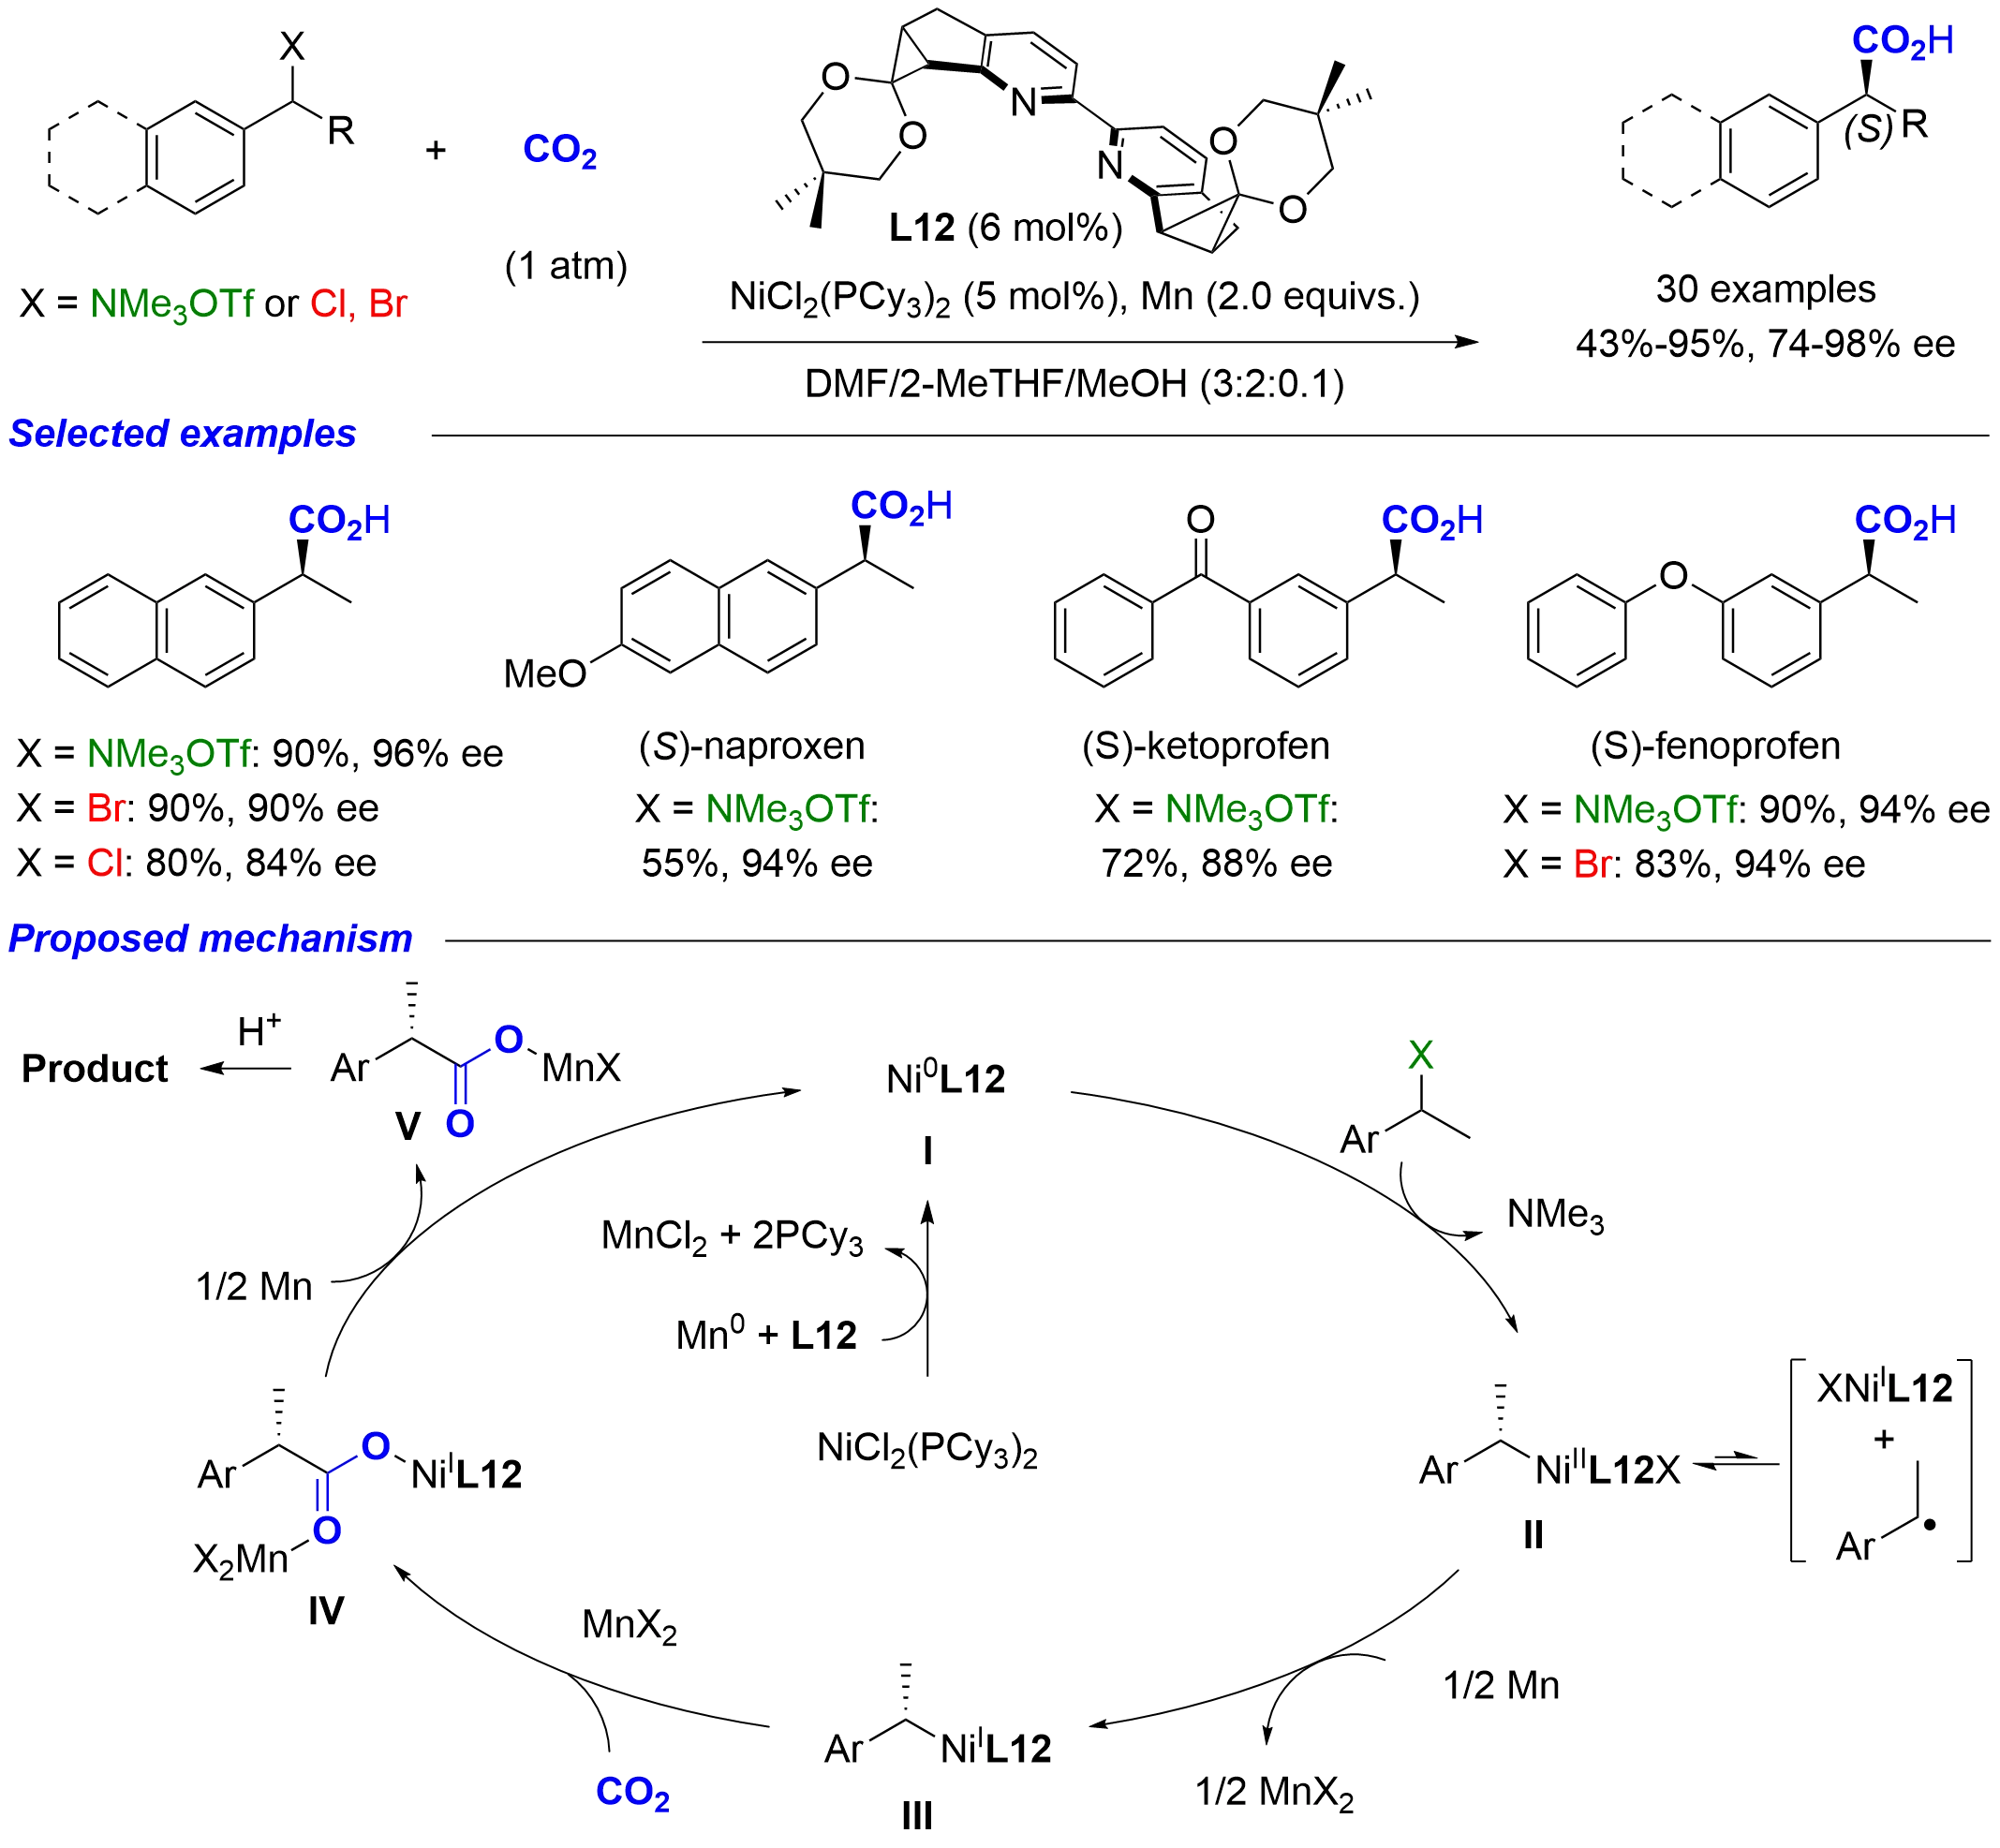
Scheme 14. Ni-catalyzed enantioconvergent carboxylation of benzylic (pseudo)halides .
The merger of asymmetric transition metal catalysis and organic electrosynthesis that employs electrons as redox reagents to generate reactive intermediates represents a noteworthy breakthrough in the research of CO2 carboxylation. In 2024, Guo, Yu and coworkers reported such a nickel-electrocatalytic asymmetric reductive carboxylation of racemic propargylic carbonates with CO2 (Scheme 15)[32]. With the combination of NiBr2 and pyridine-bisoxazoline (PYBOX) as the chiral catalyst, using an undivided cell comprising a Ni cathode and a Zn anode under constant current electrolysis at 5 mA at 0 °C and 1 atm CO2, chiral propargylic carboxylic acids could be obtained in 42-93% yield with 70-98% ee. The method accommodated diverse aryl and alkyl substituents as well as complex drug derivatives, and was applied to the asymmetric total synthesis of (S)-arundic acid, (R)-PIA, (S)-chizhine D, (S)-cochlearin G, and (S, S)-alexidine. It was proposed that the Ni(II) pre-catalyst is first reduced at the cathode to generate the active Ni(0) species I. Oxidative addition of I with the propargylic carbonate affords the π-propargyl Ni(II) intermediate II. A dynamic equilibrium is established between intermediate II and an propargyl radical species, which is key to the enantioconvergent transformation. Subsequently, II undergoes a second reduction at the cathode to furnish the η1-allenyl Ni(I) species III, whose reactivity switches from electrophilic to nucleophilic. Under the stereochemical induction of the chiral PYBOX ligand, III undergoes stereoselective nucleophilic addition to CO2 to afford the chiral nickel carboxylate IV. Protonation of IV releases the target product, while the resulting Ni(I) species is re-reduced to Ni(0) species I at the cathode to complete the catalytic cycle.
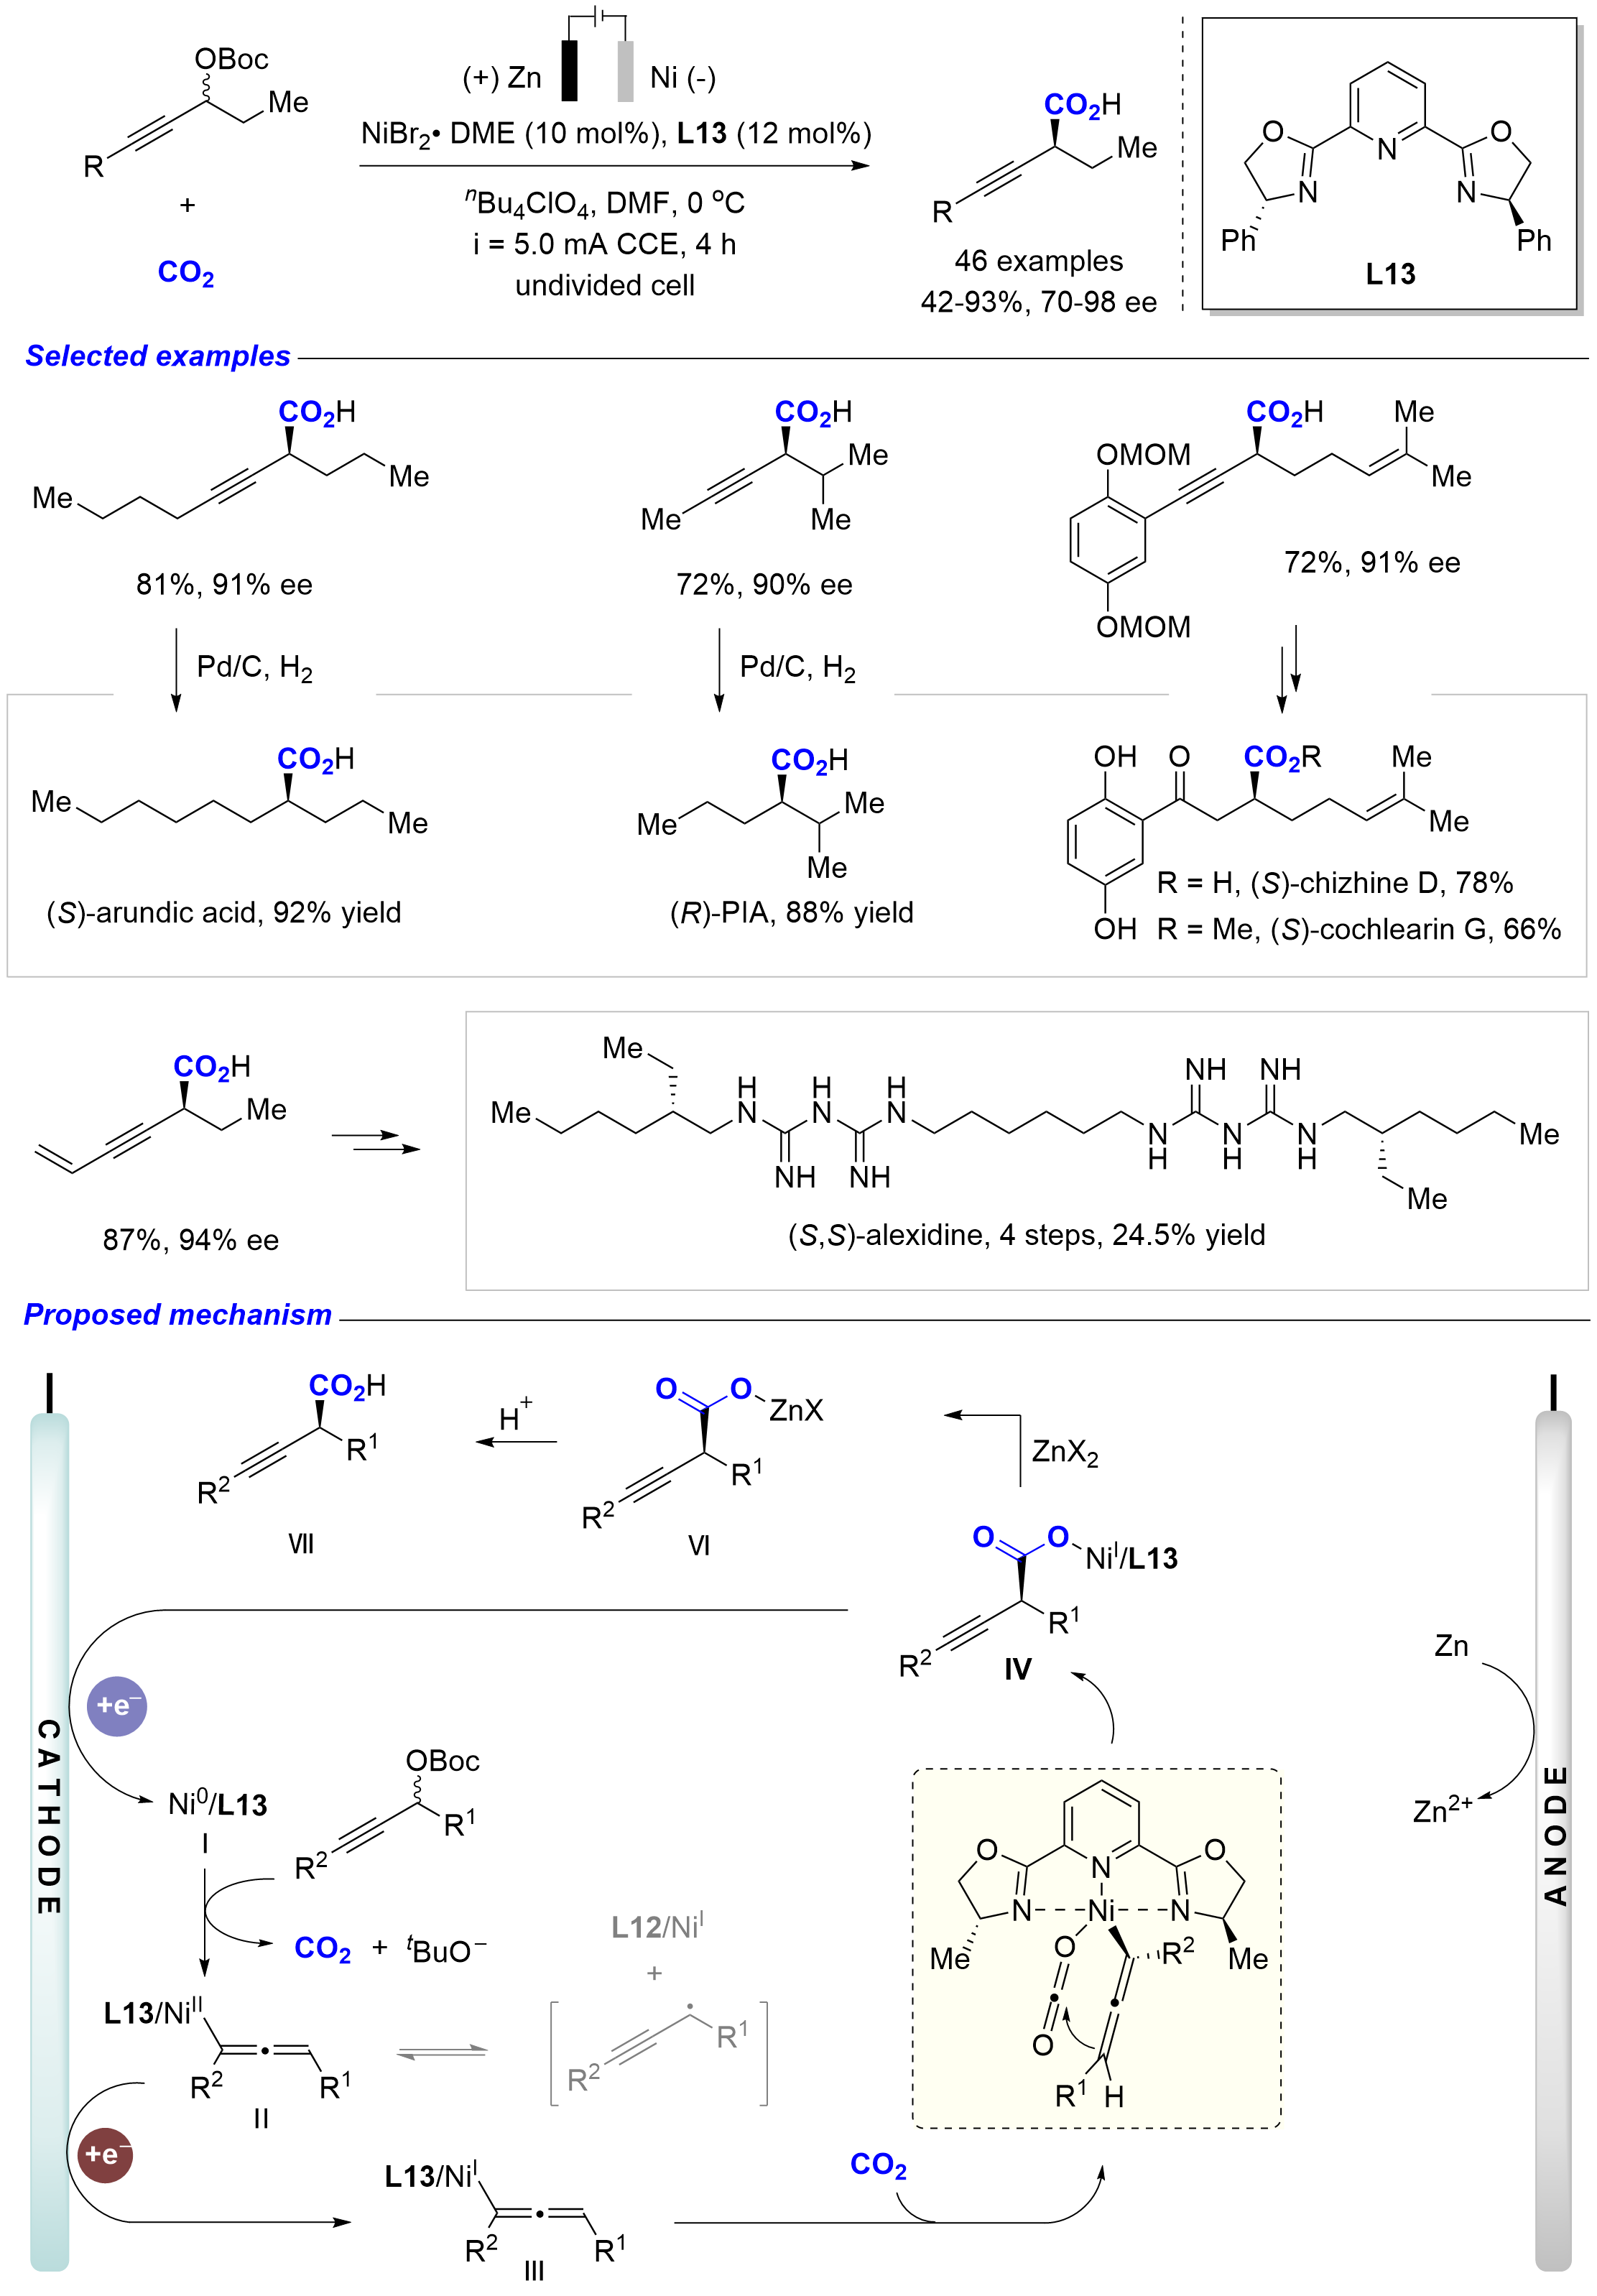
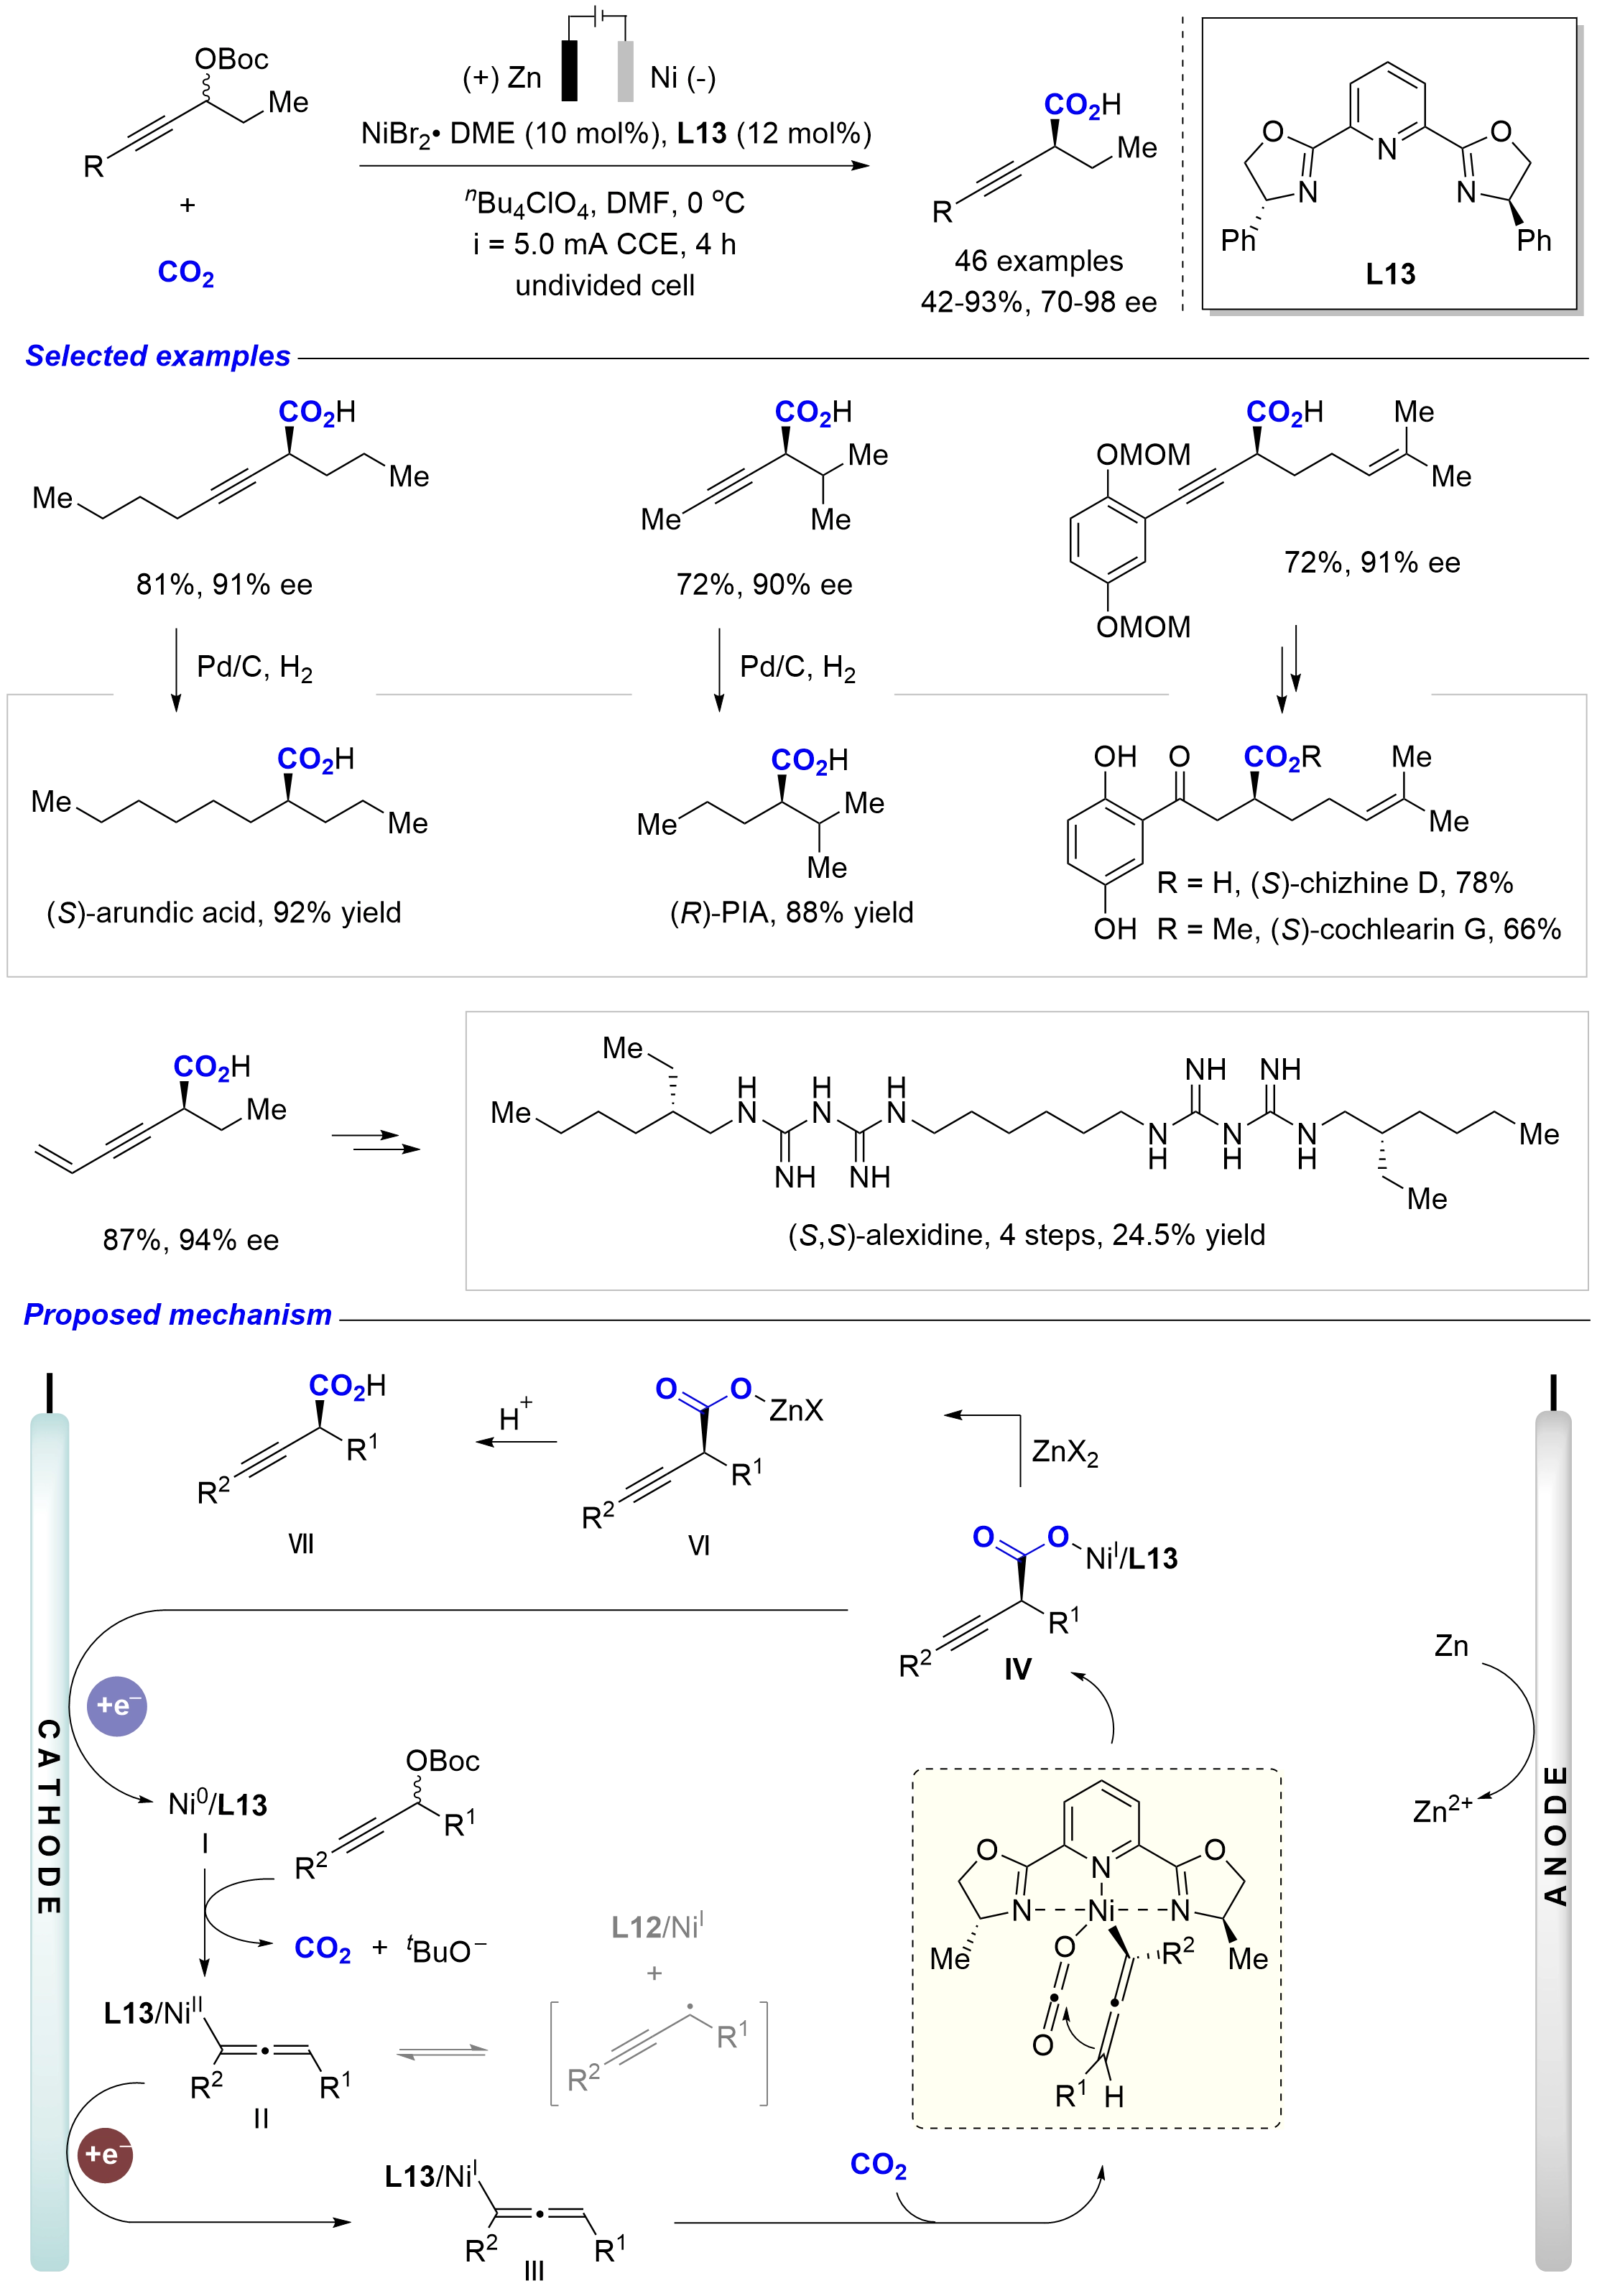
Scheme 15. Ni-electrocatalytic asymmetric reductive carboxylation of propargylic carbonates.
Notably, the metal-catalyzed carboxylation of propargylic esters with CO2 could not only yield propargylic carboxylic acids but also afford 2,3-allenoic acids in a regiodivergent manner. Zhou and coworkers found that by using the bidentate bipyridine as the ligand, the nickel-catalyzed electro-carboxylation of propargylic esters with CO2 under a constant current of 20 mA with carbon paper as the cathode and a Mg plate as the anode could produce mono-, di-, tri-, and tetra-substituted 2,3-allenoic acids with a broad substrate scope under mild conditions[33].
In addition to research on the construction of centrally chiral carboxylic acids, the catalytic enantioselective construction of axially chiral carboxylic acids with CO2 has also been explored. In 2024, Yu, Gui and coworkers pioneered a nickel-catalyzed dynamic kinetic asymmetric reductive carboxylation of azaheteroaryl triflates with CO2 for the construction of axially chiral carboxylic acids (Scheme 16)[34]. Under 1 atm CO2, with NiBr2·DME as the catalyst precursor, a chiral isoquinoline-oxazoline as the ligand, and Zn powder as the reductant, the reaction was conducted in DMSO at room temperature using LiOtBu and MgCl2 as additives. The target carboxylic acids were obtained in moderate to good yields with up to 92% ee. Notably, this research represents the first catalytic asymmetric construction of axially chiral carboxylic acids with CO2. Mechanistic studies suggested that the nitrogen atom of the isoquinoline coordinates with the in situ generated chiral Ni(0)L14 complex, facilitating oxidative addition of the C-O bond in the substrate to generate five-membered cationic nickelacyclic Ar-Ni(II)L14 diastereomers III. Single-electron reduction of this intermediate by zinc powder affords the more reactive Ar-Ni(I)L14 species IV. The strong coordination of the nitrogen atom to nickel increases the dihedral angle of the axial system, allowing epimerization of both the Ar-Ni(II)L14 and Ar-Ni(I)L14 species, thereby enabling enantioconvergent conversion of the racemic substrate. Subsequently, IV undergoes an enantioselective electrophilic reaction with CO2, followed by reductive elimination to furnish the axially chiral carboxylic acid product and regenerate the active Ni(0) species. It was proposed that the tert-butoxide base could increase the CO2 concentration in solution, thus suppressing the undesired reductive protonation. The MgCl2 additive could activate CO2 as a Lewis acid and facilitate the migratory insertion by stabilizing the ArCO2Ni(I) intermediate, thus increasing the chemoselectivity.
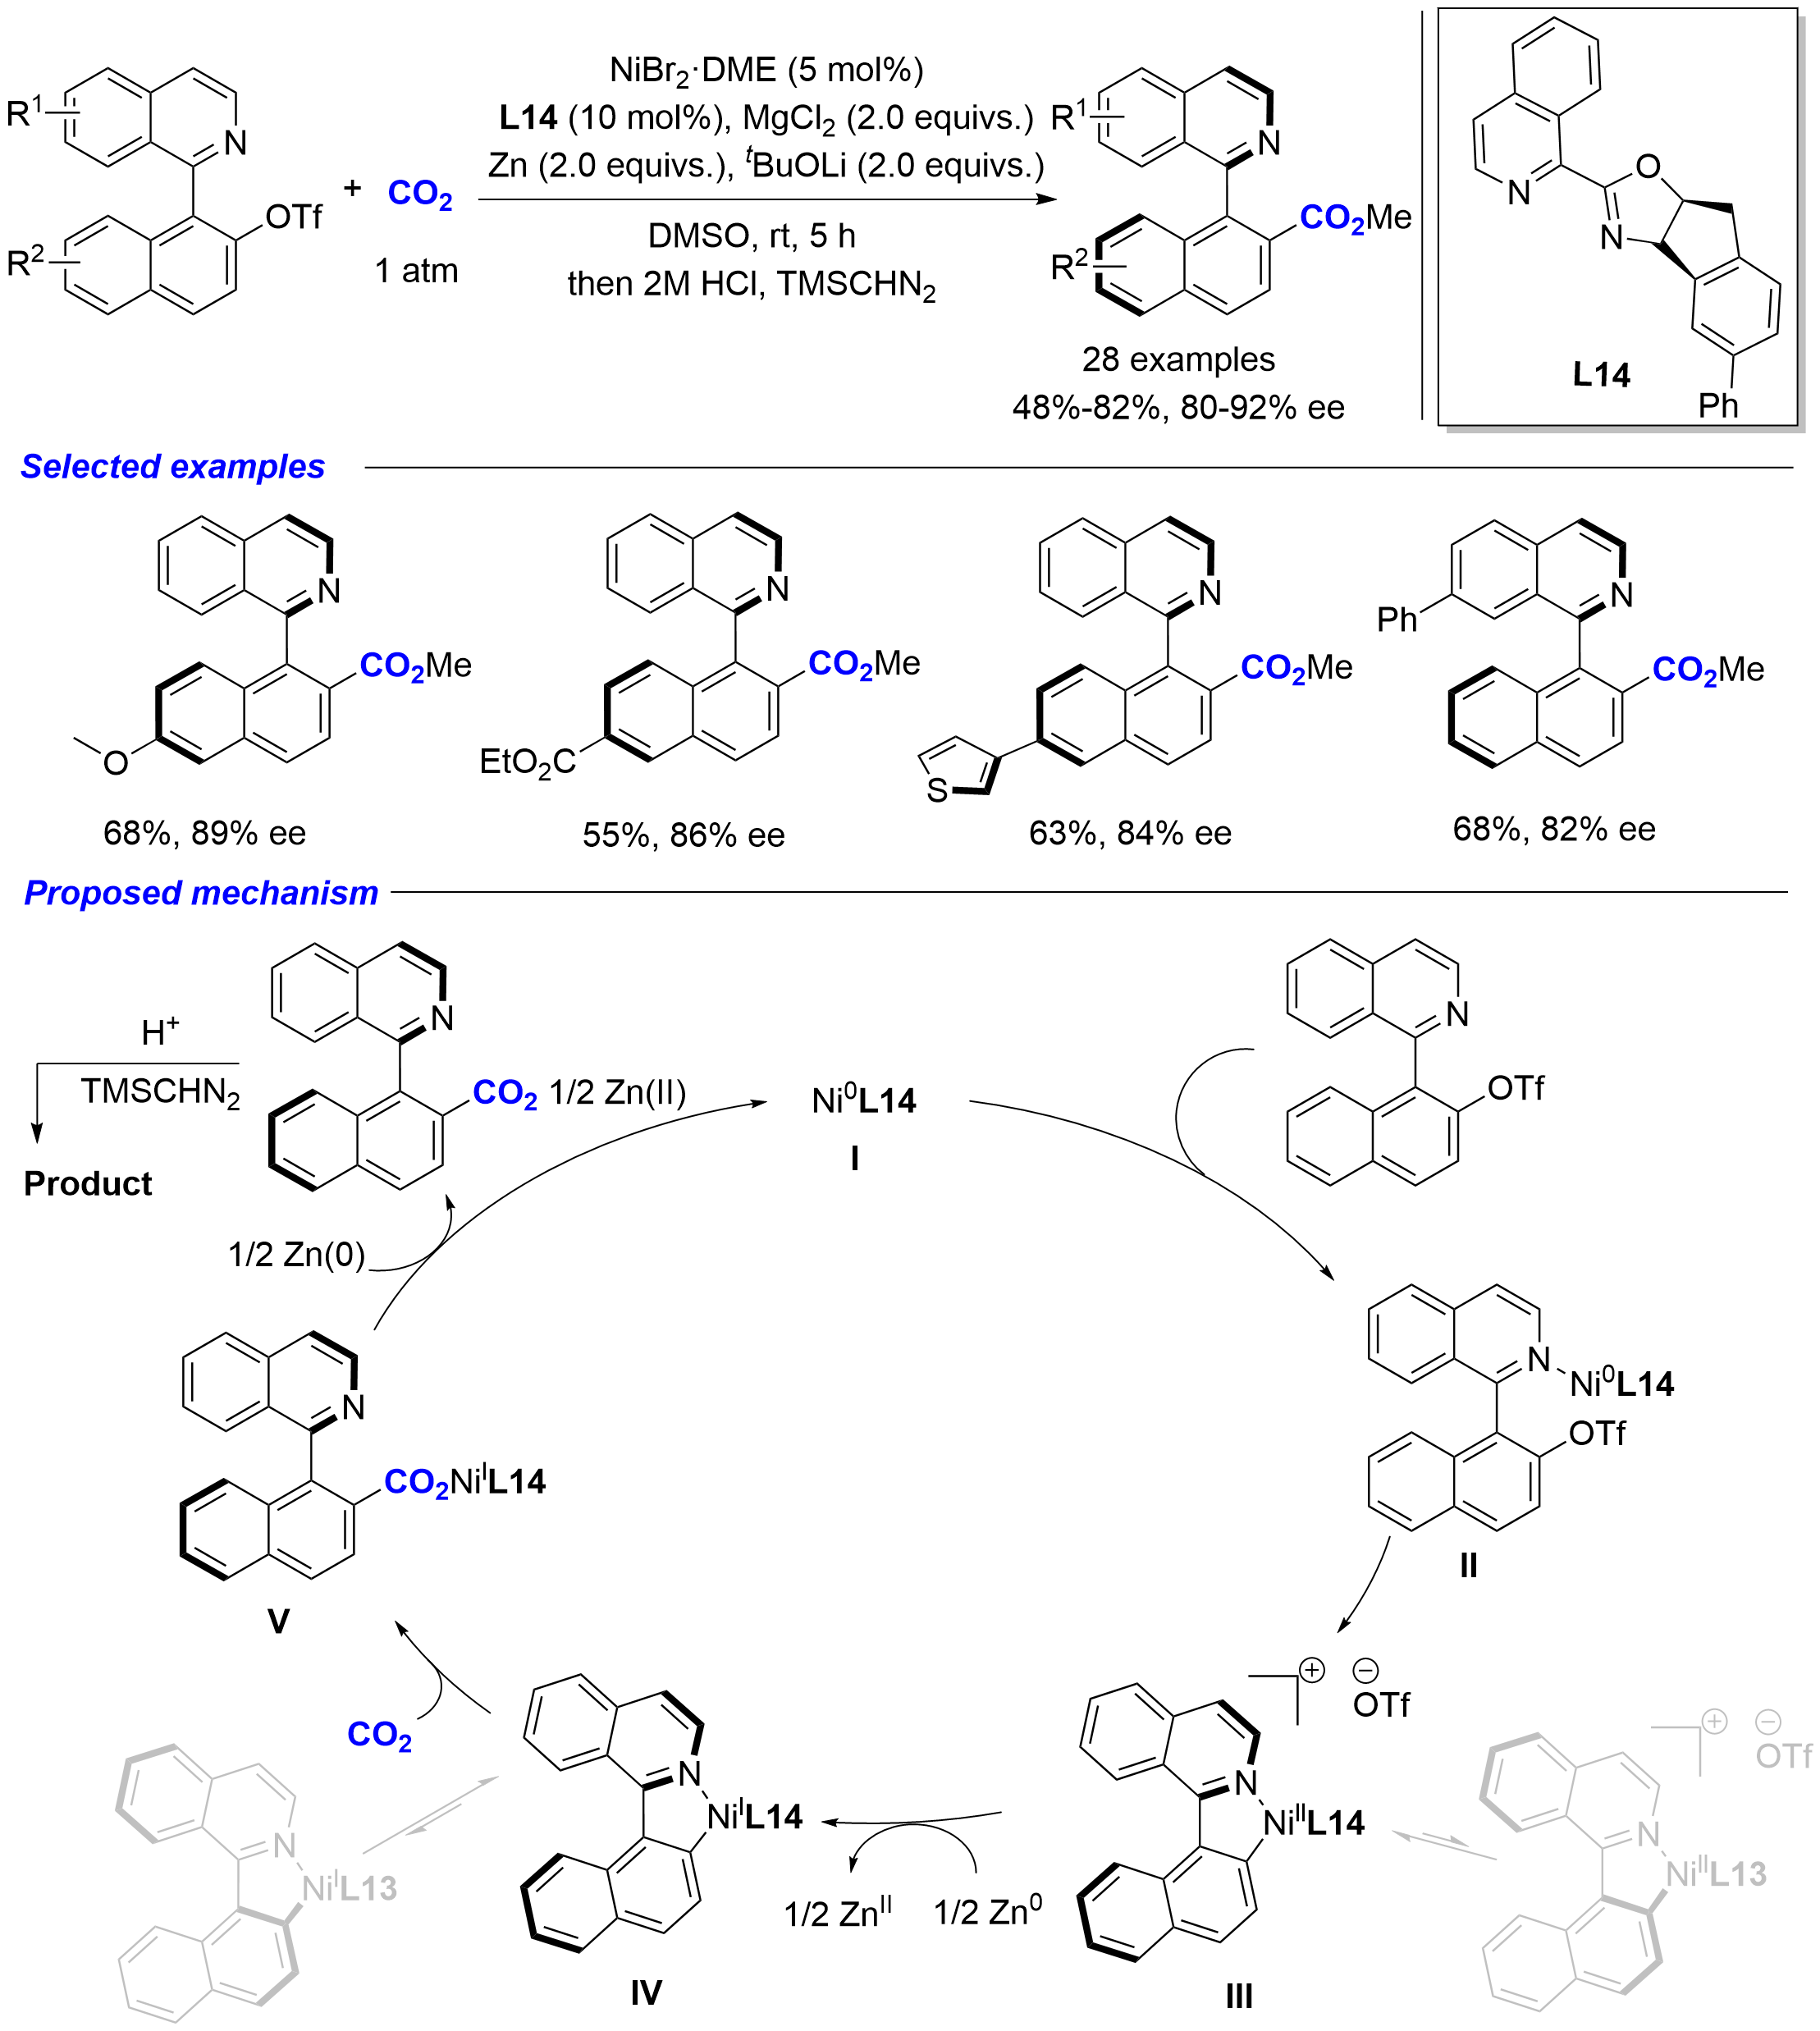
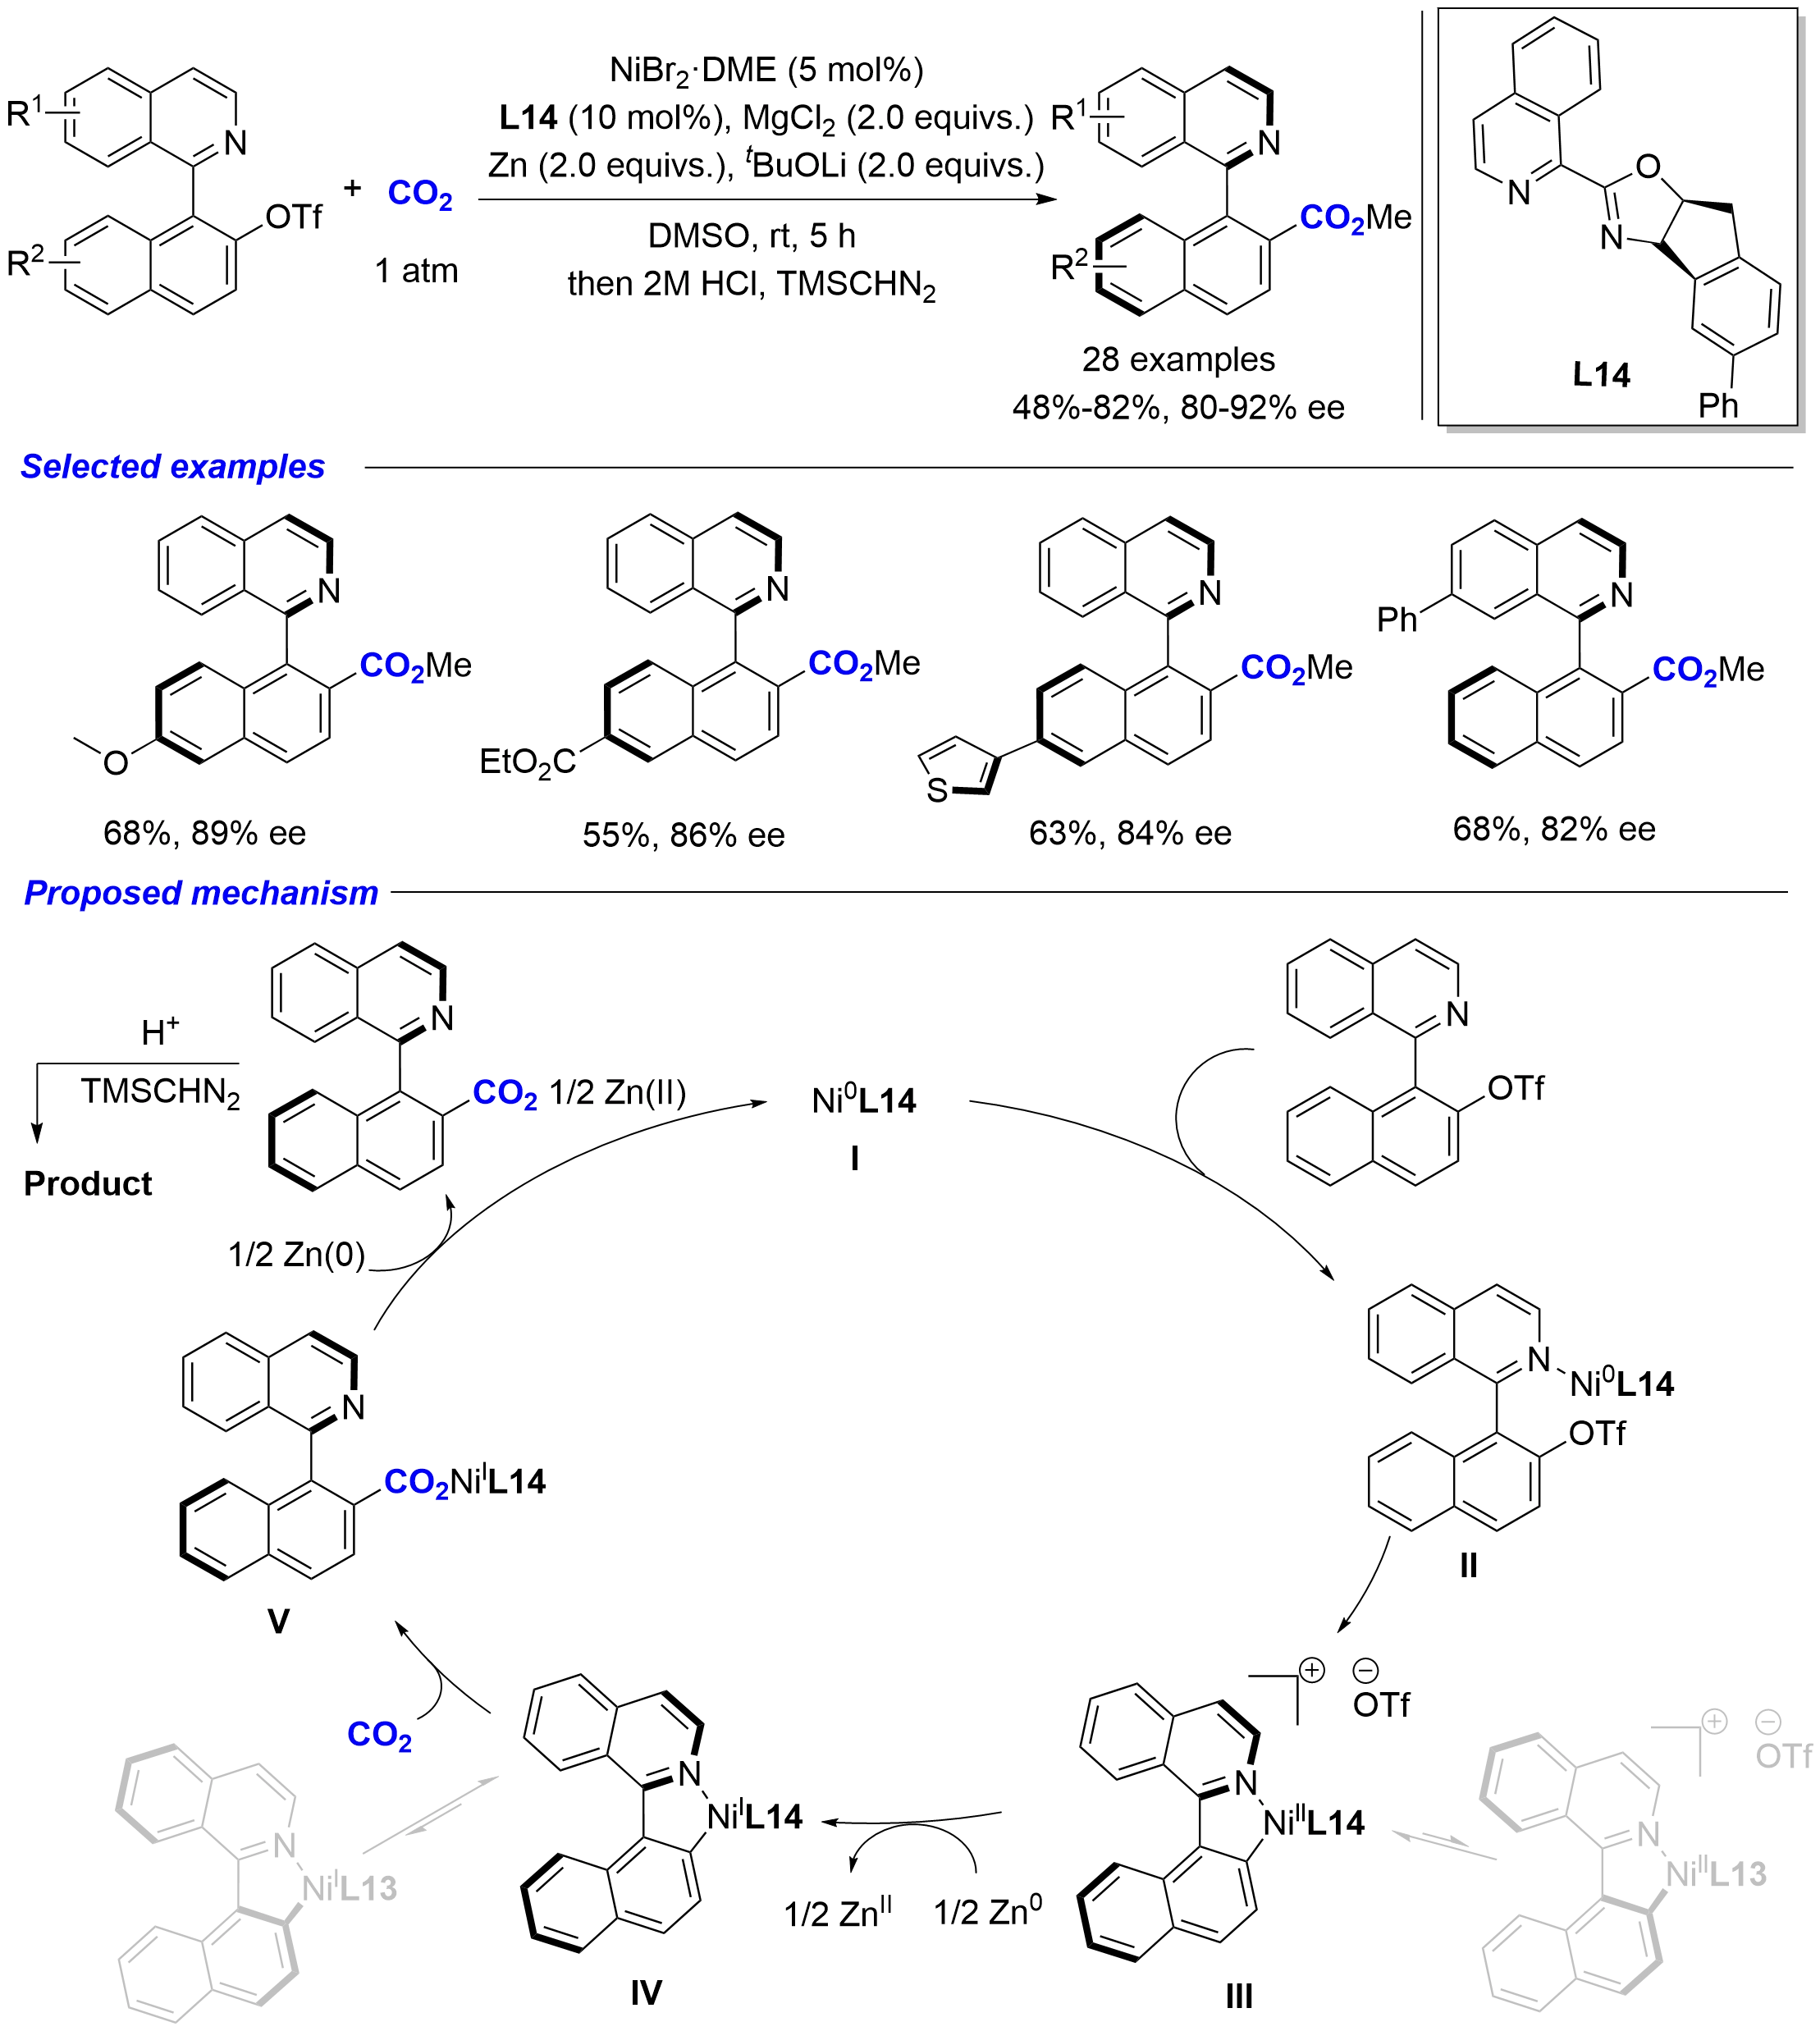
Scheme 16. Ni-catalyzed dynamic kinetic asymmetric reductive carboxylation to axially chiral carboxylic acids.
In 2025, Liu and coworkers developed an enantioconvergent carboxylation-alkylation reaction of racemic heterobiaryl (pseudo)halides with CO2 and alkyl bromides via an elegant synergistic photoredox/palladium cooperative catalysis, affording axially chiral esters with high efficiency and enantioselectivity (Scheme 17)[35]. Using Pd(acac)2 as the metal precursor, an axially chiral bisphosphine ligand as the chiral source, 4CzIPN as the photocatalyst, DIPEA as the reductant, LiNTf2 as the additive, and Cs2CO3 as the base, the reaction proceeded at room temperature under blue LED irradiation. A range of alkyl bromides and heterobiaryl bromides/triflates participated smoothly, delivering the corresponding axially chiral esters in 47-92% yield with 90-96% ee. This work represents the first example of metallaphotoredox catalyzed asymmetric transformation of CO2 to axially chiral esters. Mechanistic studies revealed that the Pd(0)/L15 species undergoes oxidative addition with the racemic heterobiaryl bromide to generate cyclopalladium intermediates I. Upon visible-light excitation, the photocatalyst 4CzIPN* reduces the cyclopalladium species I to ArPd(I)/L15 species II. The epimerization of intermediates I or II results in a dynamic kinetic asymmetric transformation. Subsequent CO2 coordination and migratory insertion afford Pd(I) carboxylate III, which is trapped by the alkyl bromide via an SN2 pathway to furnish the axially chiral ester and the released Pd(I)L15 species is reduced to Pd(0)L15 to close the catalytic cycle. The in situ trapping of the chiral carboxylate anion by the alkyl bromide is crucial for suppressing racemization.
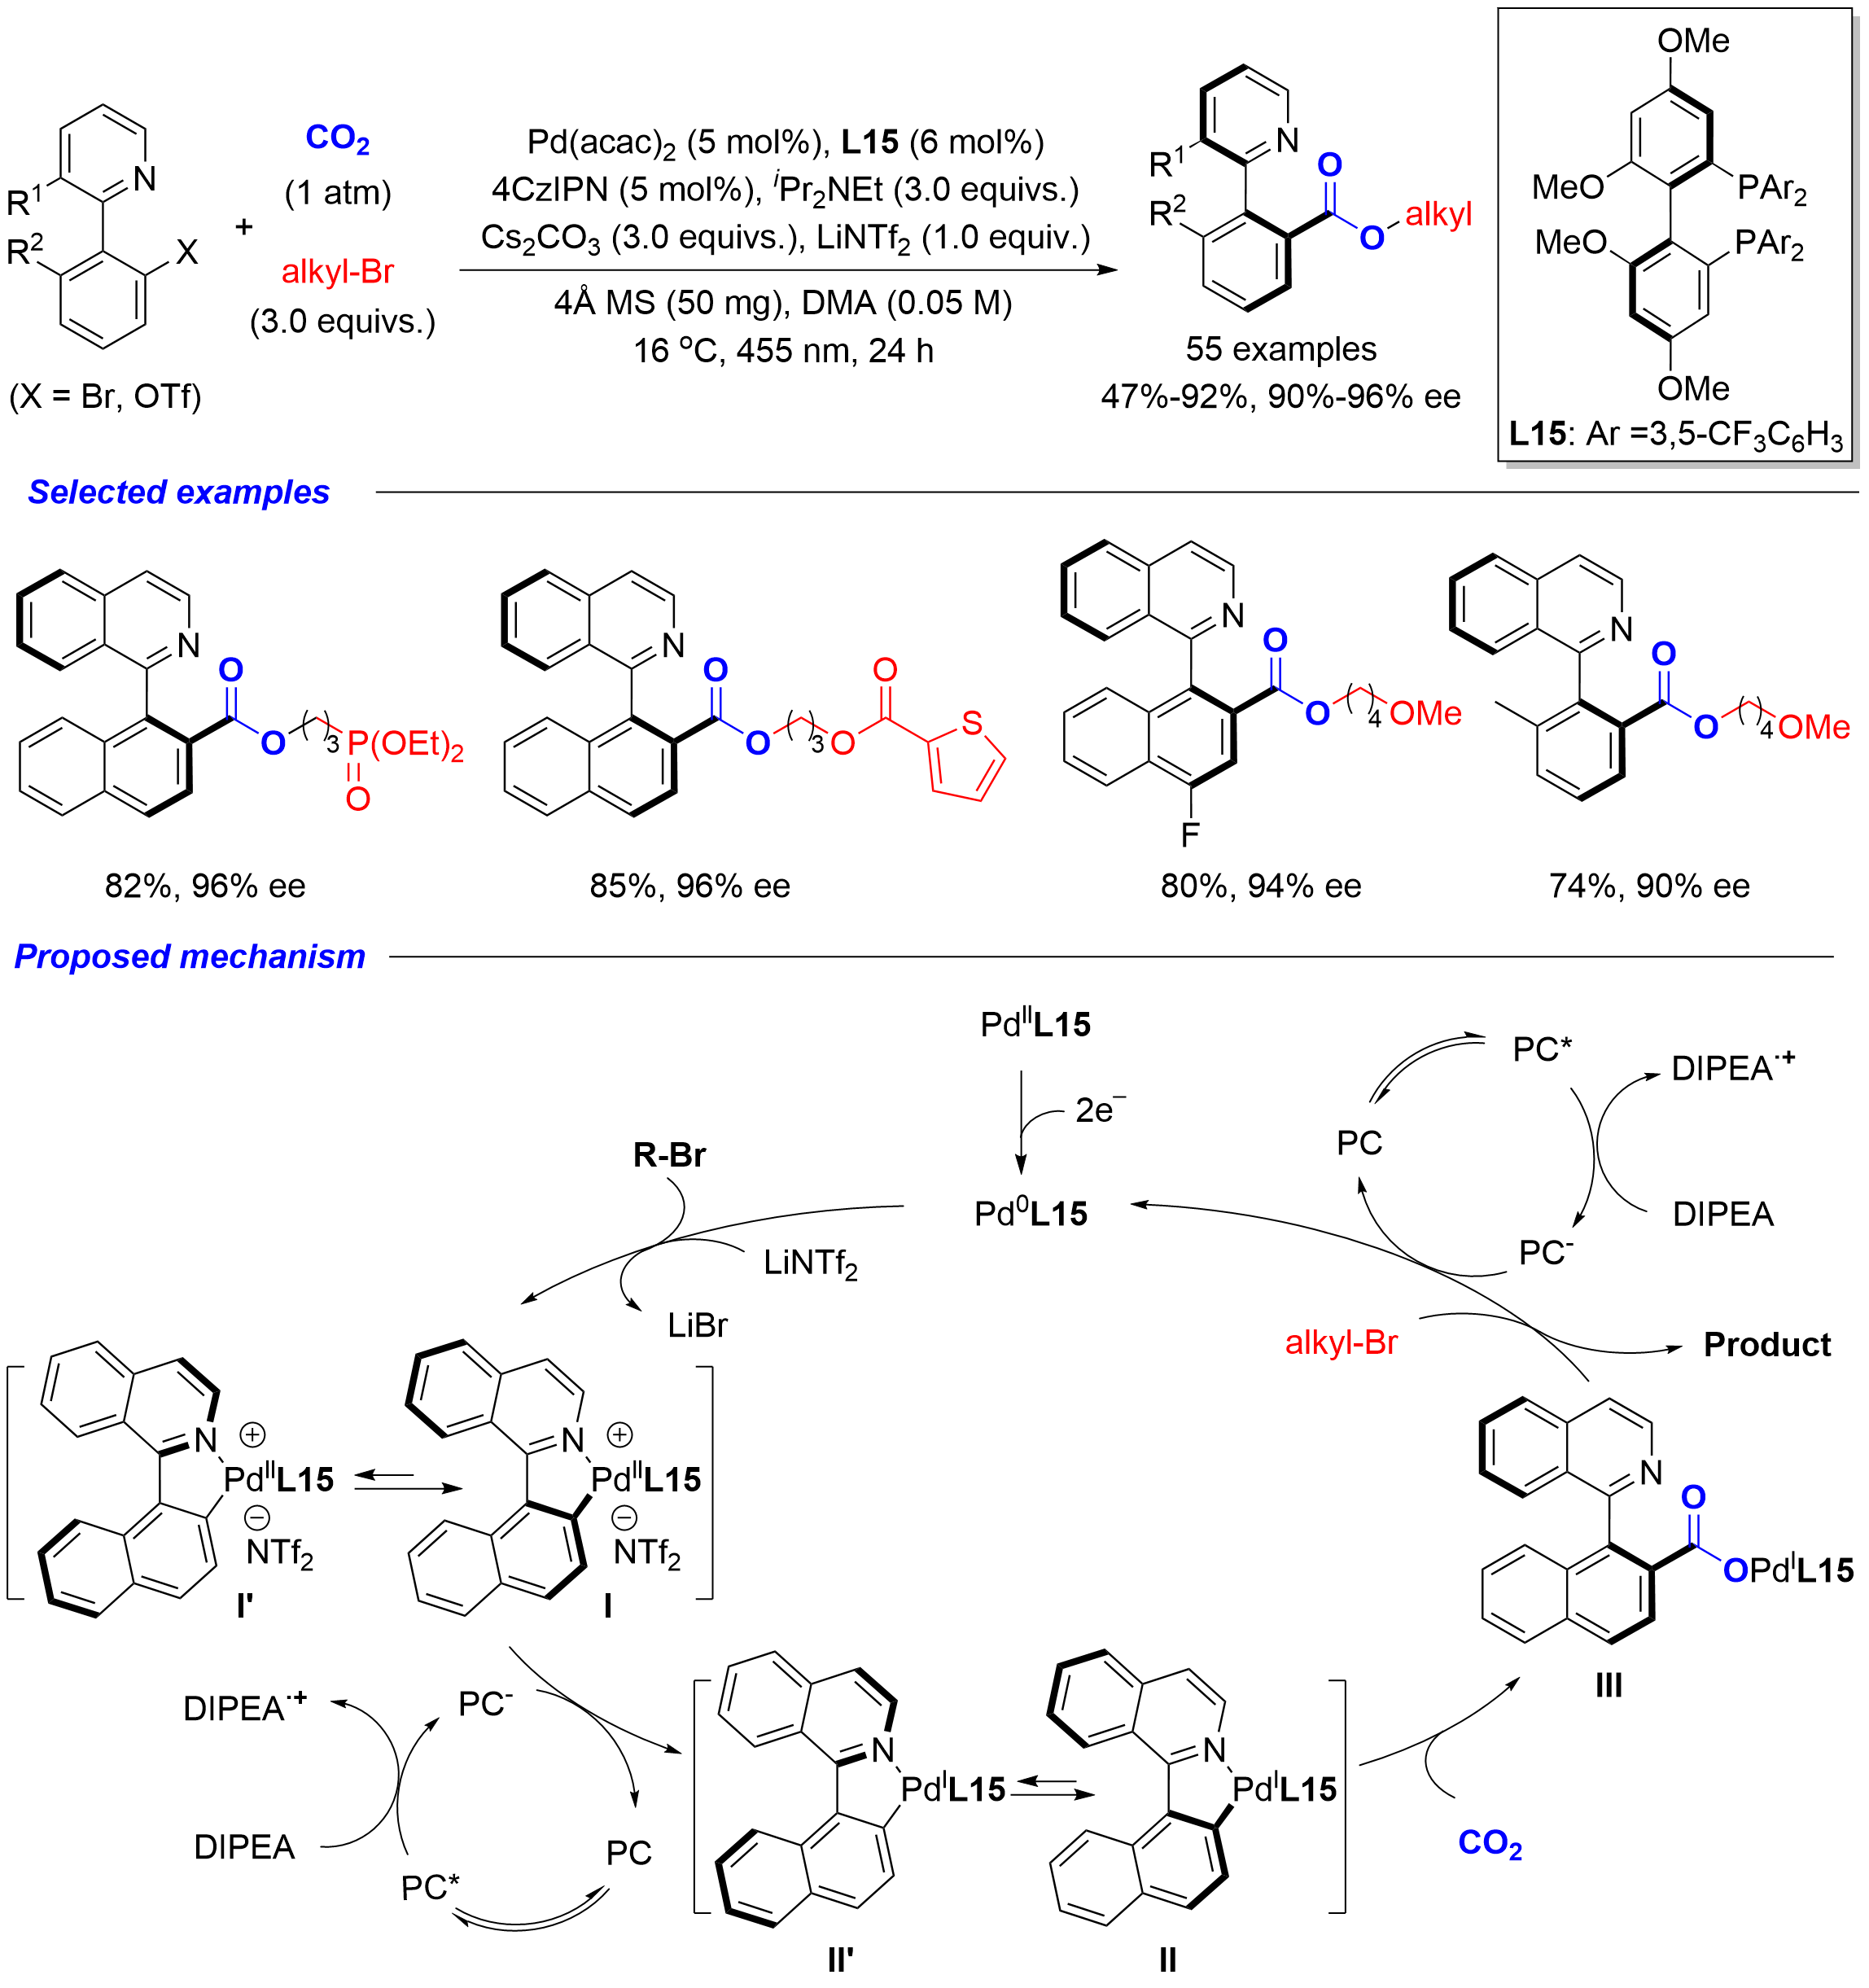
Scheme 17. Photoredox/Pd-catalyzed asymmetric carboxylation-alkylation to axially chiral esters.
Later in 2026, Liu, Gao and coworkers further reported the first metallaphotoredox-catalyzed desymmetric carboxylation of prochiral biaryl triflates with CO2 (Scheme 18)[36]. Using Pd(OAc)2/DTBM-SEGPHOS (L16) as the chiral palladium catalyst, 4DPAIPN as the organic photocatalyst, DIPEA as the reductant, and Cs2CO3 as the base in DMF at -40 °C under blue LED irradiation, axially chiral biaryl carboxylic acids were obtained in up to 83% yield with up to 96% ee. The reaction exhibited broad substrate scope, accommodating diverse aryl, naphthyl, polycyclic, and heterocyclic substituents. The resulting product could be transformed into a novel axially chiral monophosphine ligand via a three-step sequence, which demonstrated excellent performance in Pd-catalyzed asymmetric allylic alkylation and amination reactions.
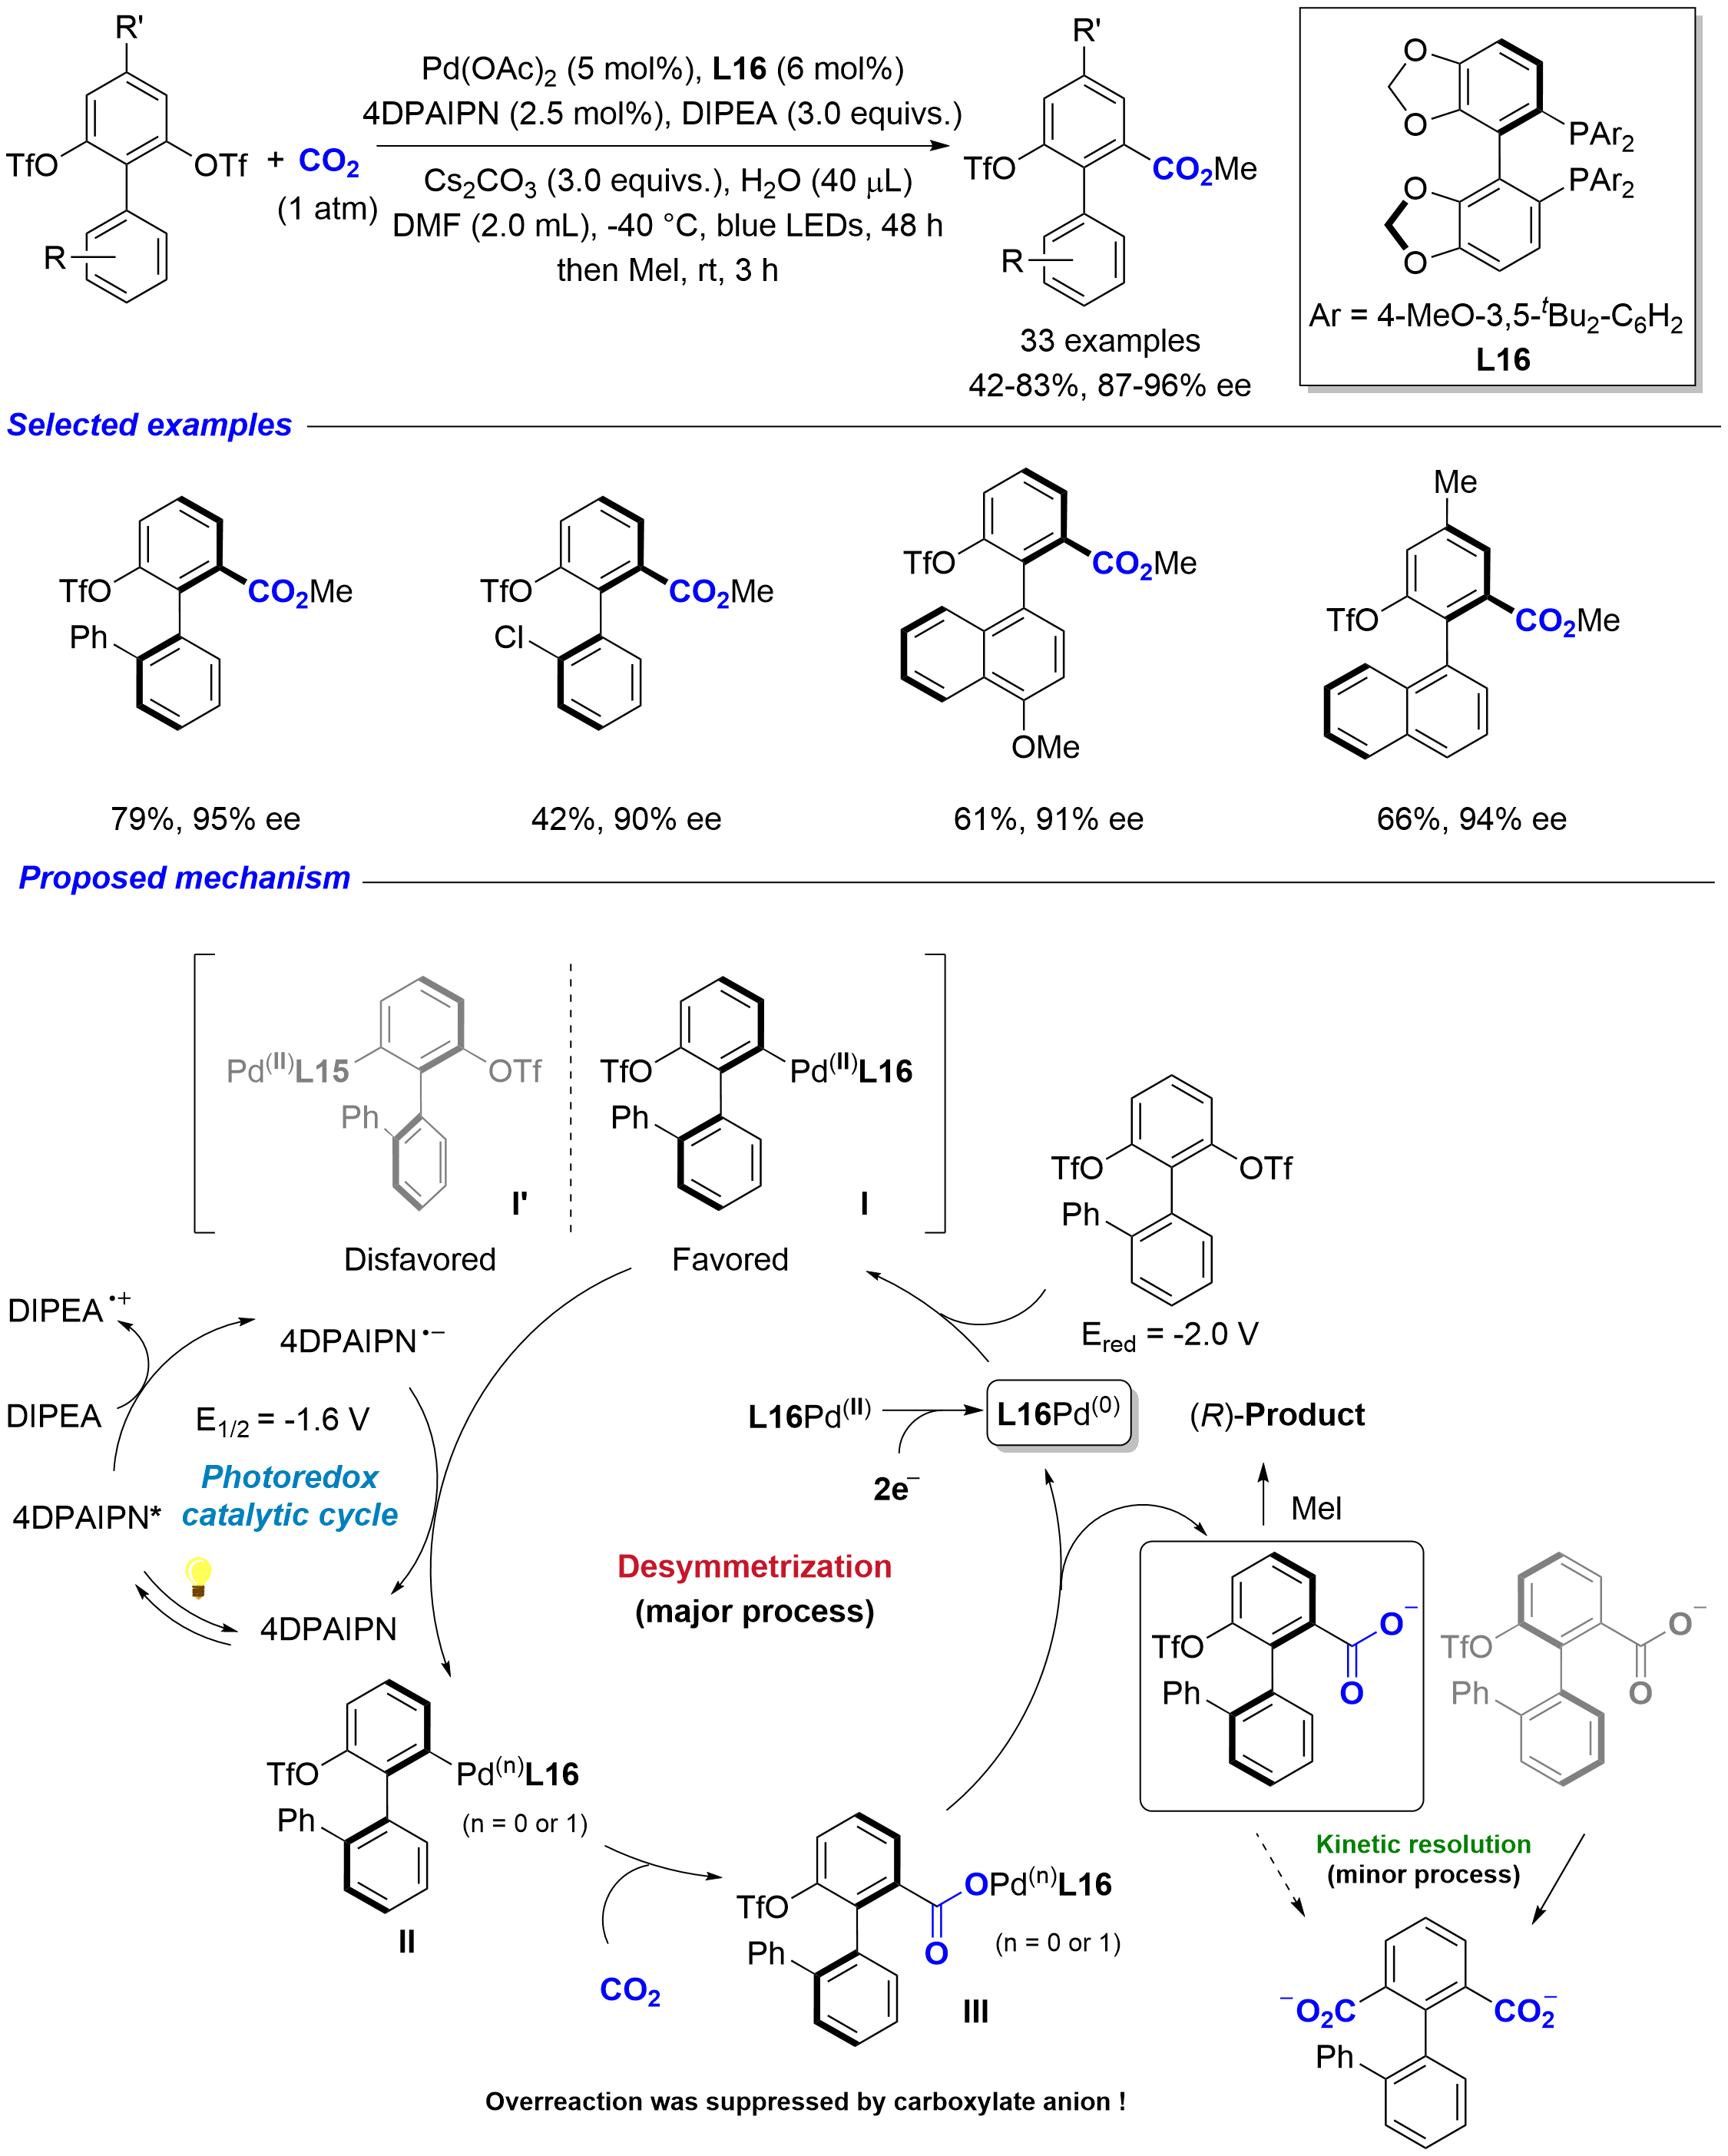
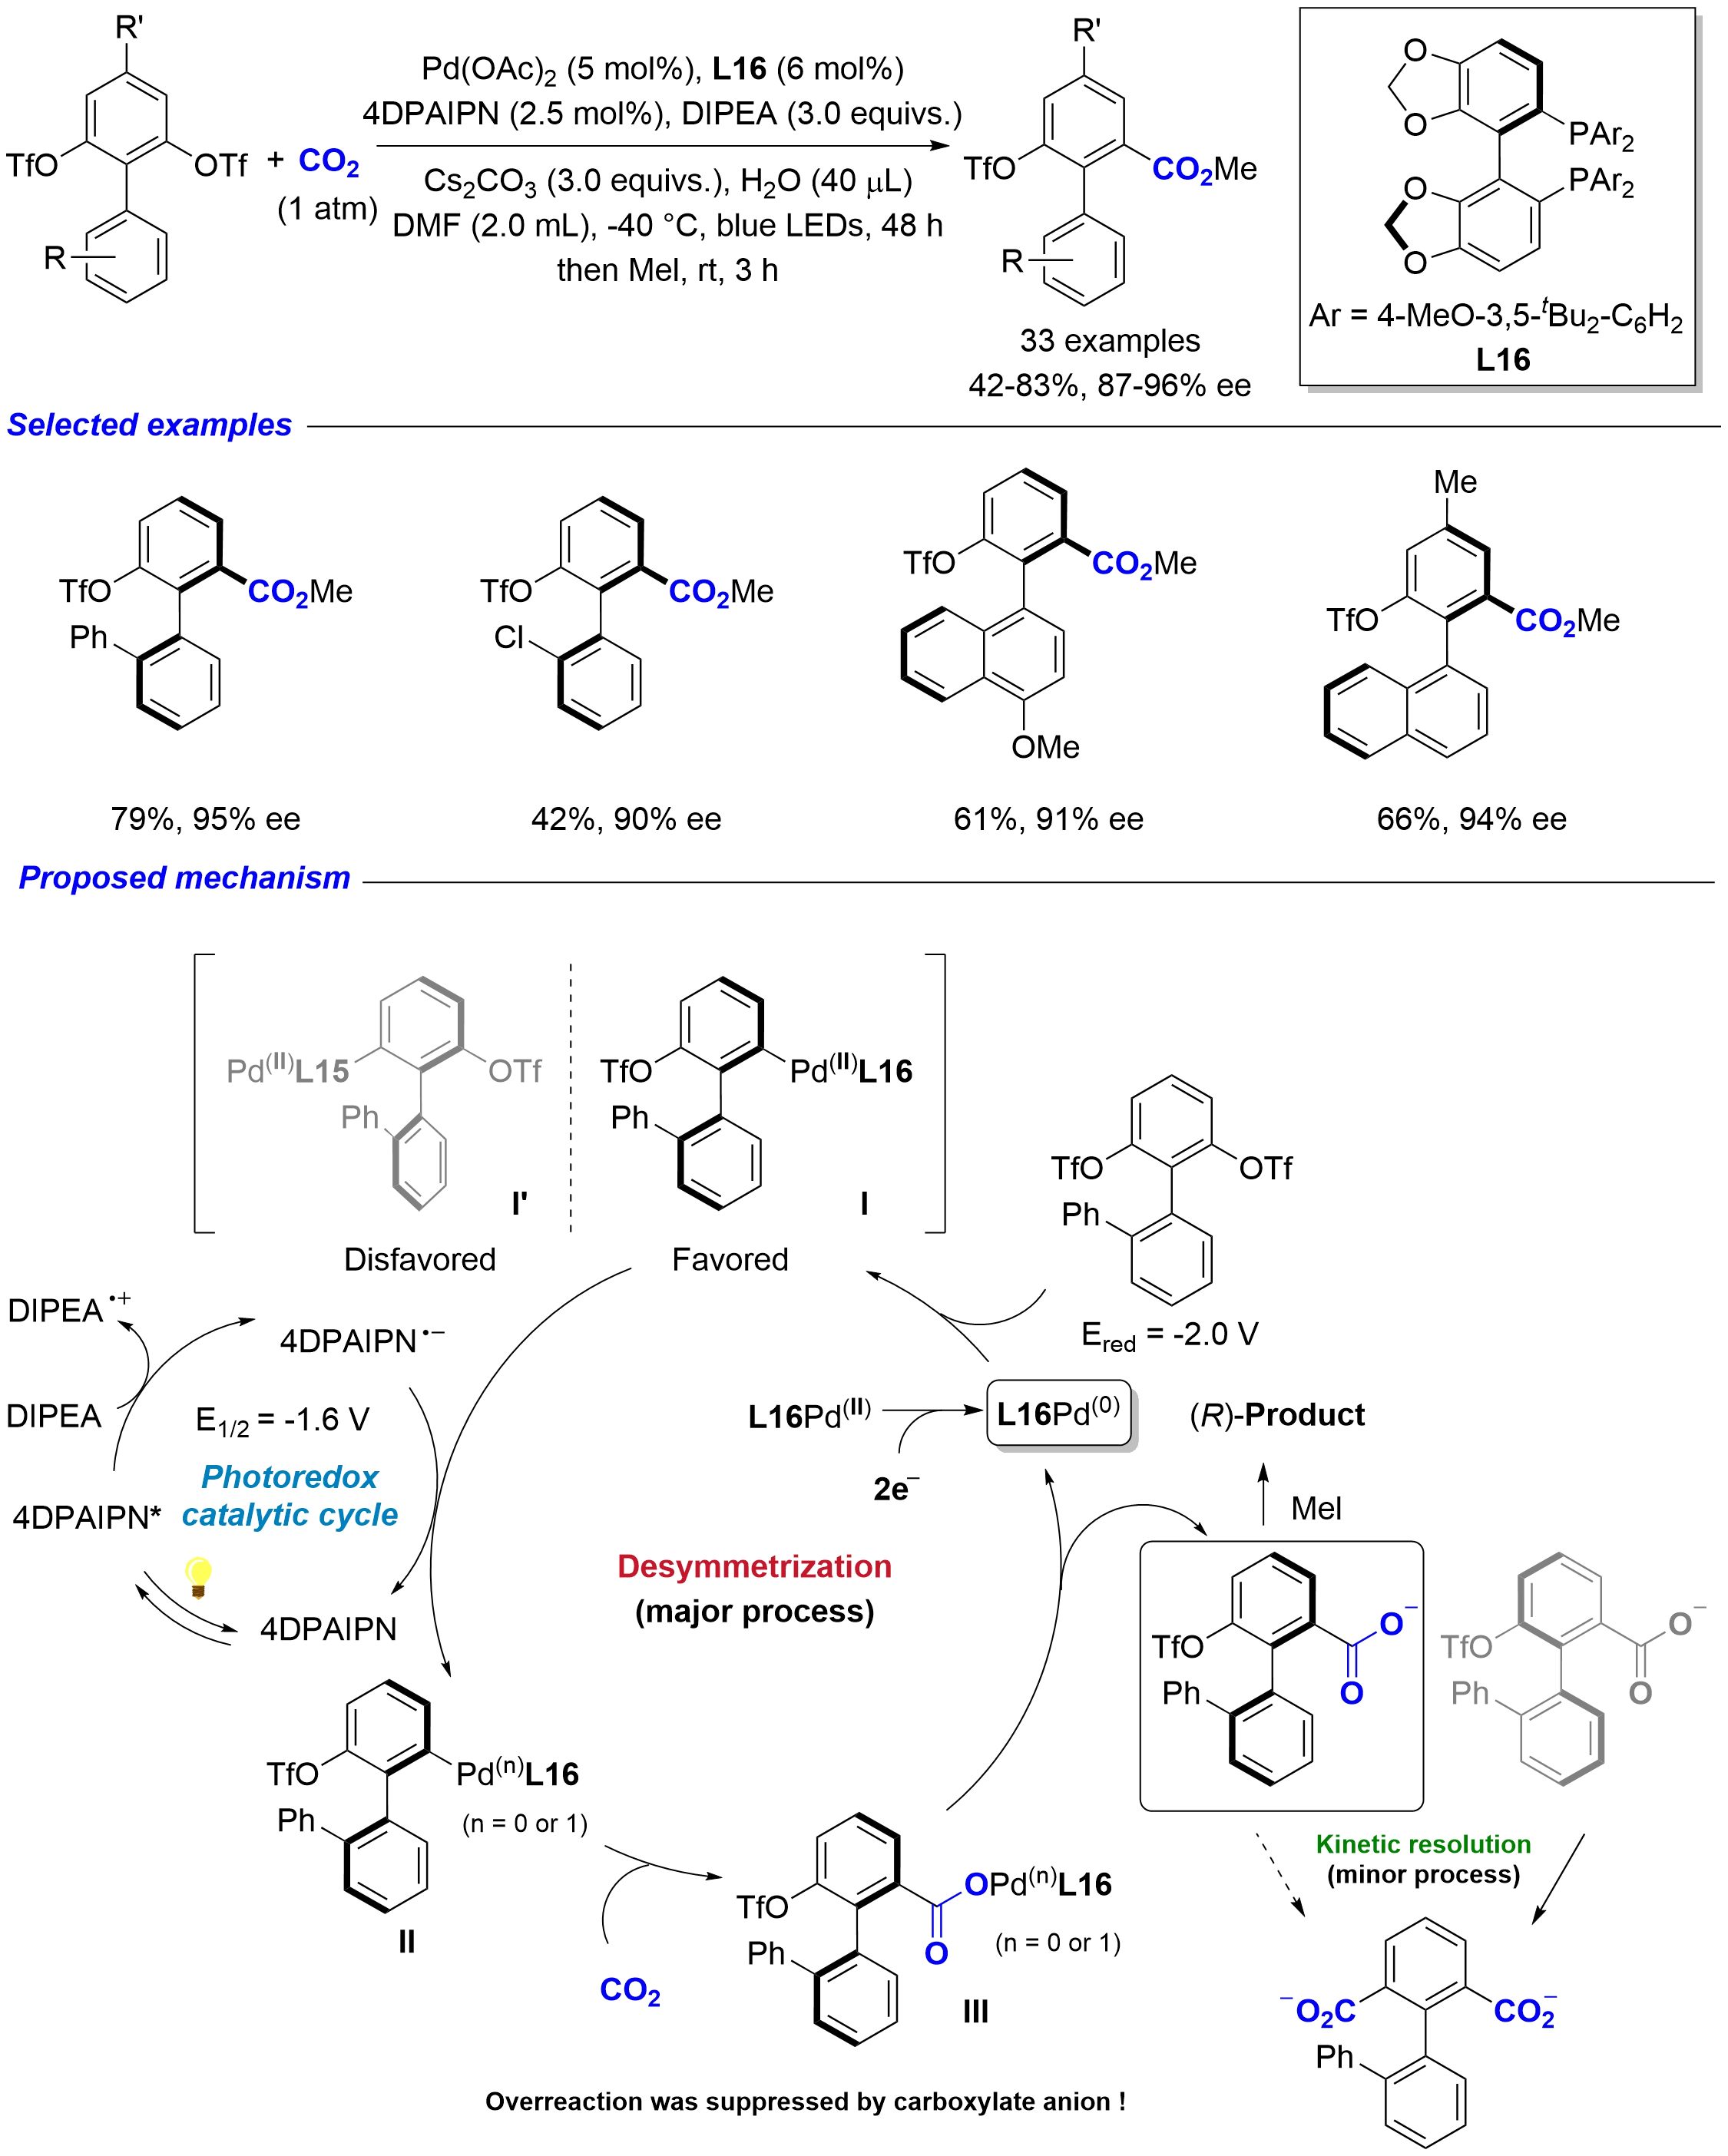
Scheme 18. Pd/photoredox-catalyzed desymmetric carboxylation of biaryl ditriflates.
Based on systematic mechanistic studies, the catalytic cycle was proposed as follows. First, the Pd(II)/L16 precatalyst is reduced to the active Pd(0)/L16 species, which then undergoes asymmetric oxidative addition with the prochiral bis(triflate) substrate to generate the ArPd(II)L16 intermediate I. Visible-light excitation of the photocatalyst 4DPAIPN* triggers single-electron transfer with DIPEA to generate the reduced species PC∙-, which effects a single-electron reduction of intermediate I to afford the ArPd(I/0)L16 species II. Subsequent coordination and migratory insertion of CO2 with II generate the palladium carboxylate intermediate III. Under basic conditions, III dissociates to release the monocarboxylate anion product while regenerating the Pd(0)/L16 catalyst. The electron-donating effect of the carboxylate anion generated from the first carboxylation, combined with the mismatched steric influence of the chiral ligand, collectively suppresses the second carboxylation, thereby ensuring high enantioselectivity and yield.
The desymmetric strategy could also be utilized for the construction of axially chiral carbamates and amides with CO2 as a C1 synthon. In 2023, Jiang, Qi and coworkers employed cyclic diaryliodonium salts as electrophiles to achieve a copper-catalyzed asymmetric three-component ring-opening coupling with CO2 and amines, enabling the synthesis of axially chiral carbamates with up to 99% ee values and good to excellent yields (Scheme 19)[37]. The reaction tolerated a broad range of cyclic and acyclic secondary and primary amines and was applied to the late-stage modification of amino acid esters, natural products, and drug molecules. Notably, by adding nBuLi to the reaction mixture to trigger an anionic Fries rearrangement, the same intermediate could be converted into axially chiral amides with retained enantioselectivity. Mechanistic studies revealed that diethylamine reduces Cu(OTf)2 in situ to generate the active Cu(I) species. This Cu(I) species coordinates with chiral ligand L17 to form the active complex I. Complex I preferentially interacts with the cyclic diaryliodonium salt via the sterically less hindered mode II (in contrast, mode II’ suffers from significant steric repulsion between the methyl group of the iodonium salt and the phenyl group of L17), followed by oxidative addition to afford the axially chiral Cu(III) intermediate III. Concurrently, CO2 reacts with the amine in situ to generate the carbamate anion IV, which coordinates to the Cu(III) center to furnish intermediate V. Finally, intermediate V undergoes reductive elimination to release the product and regenerate the active Cu(I) species I.
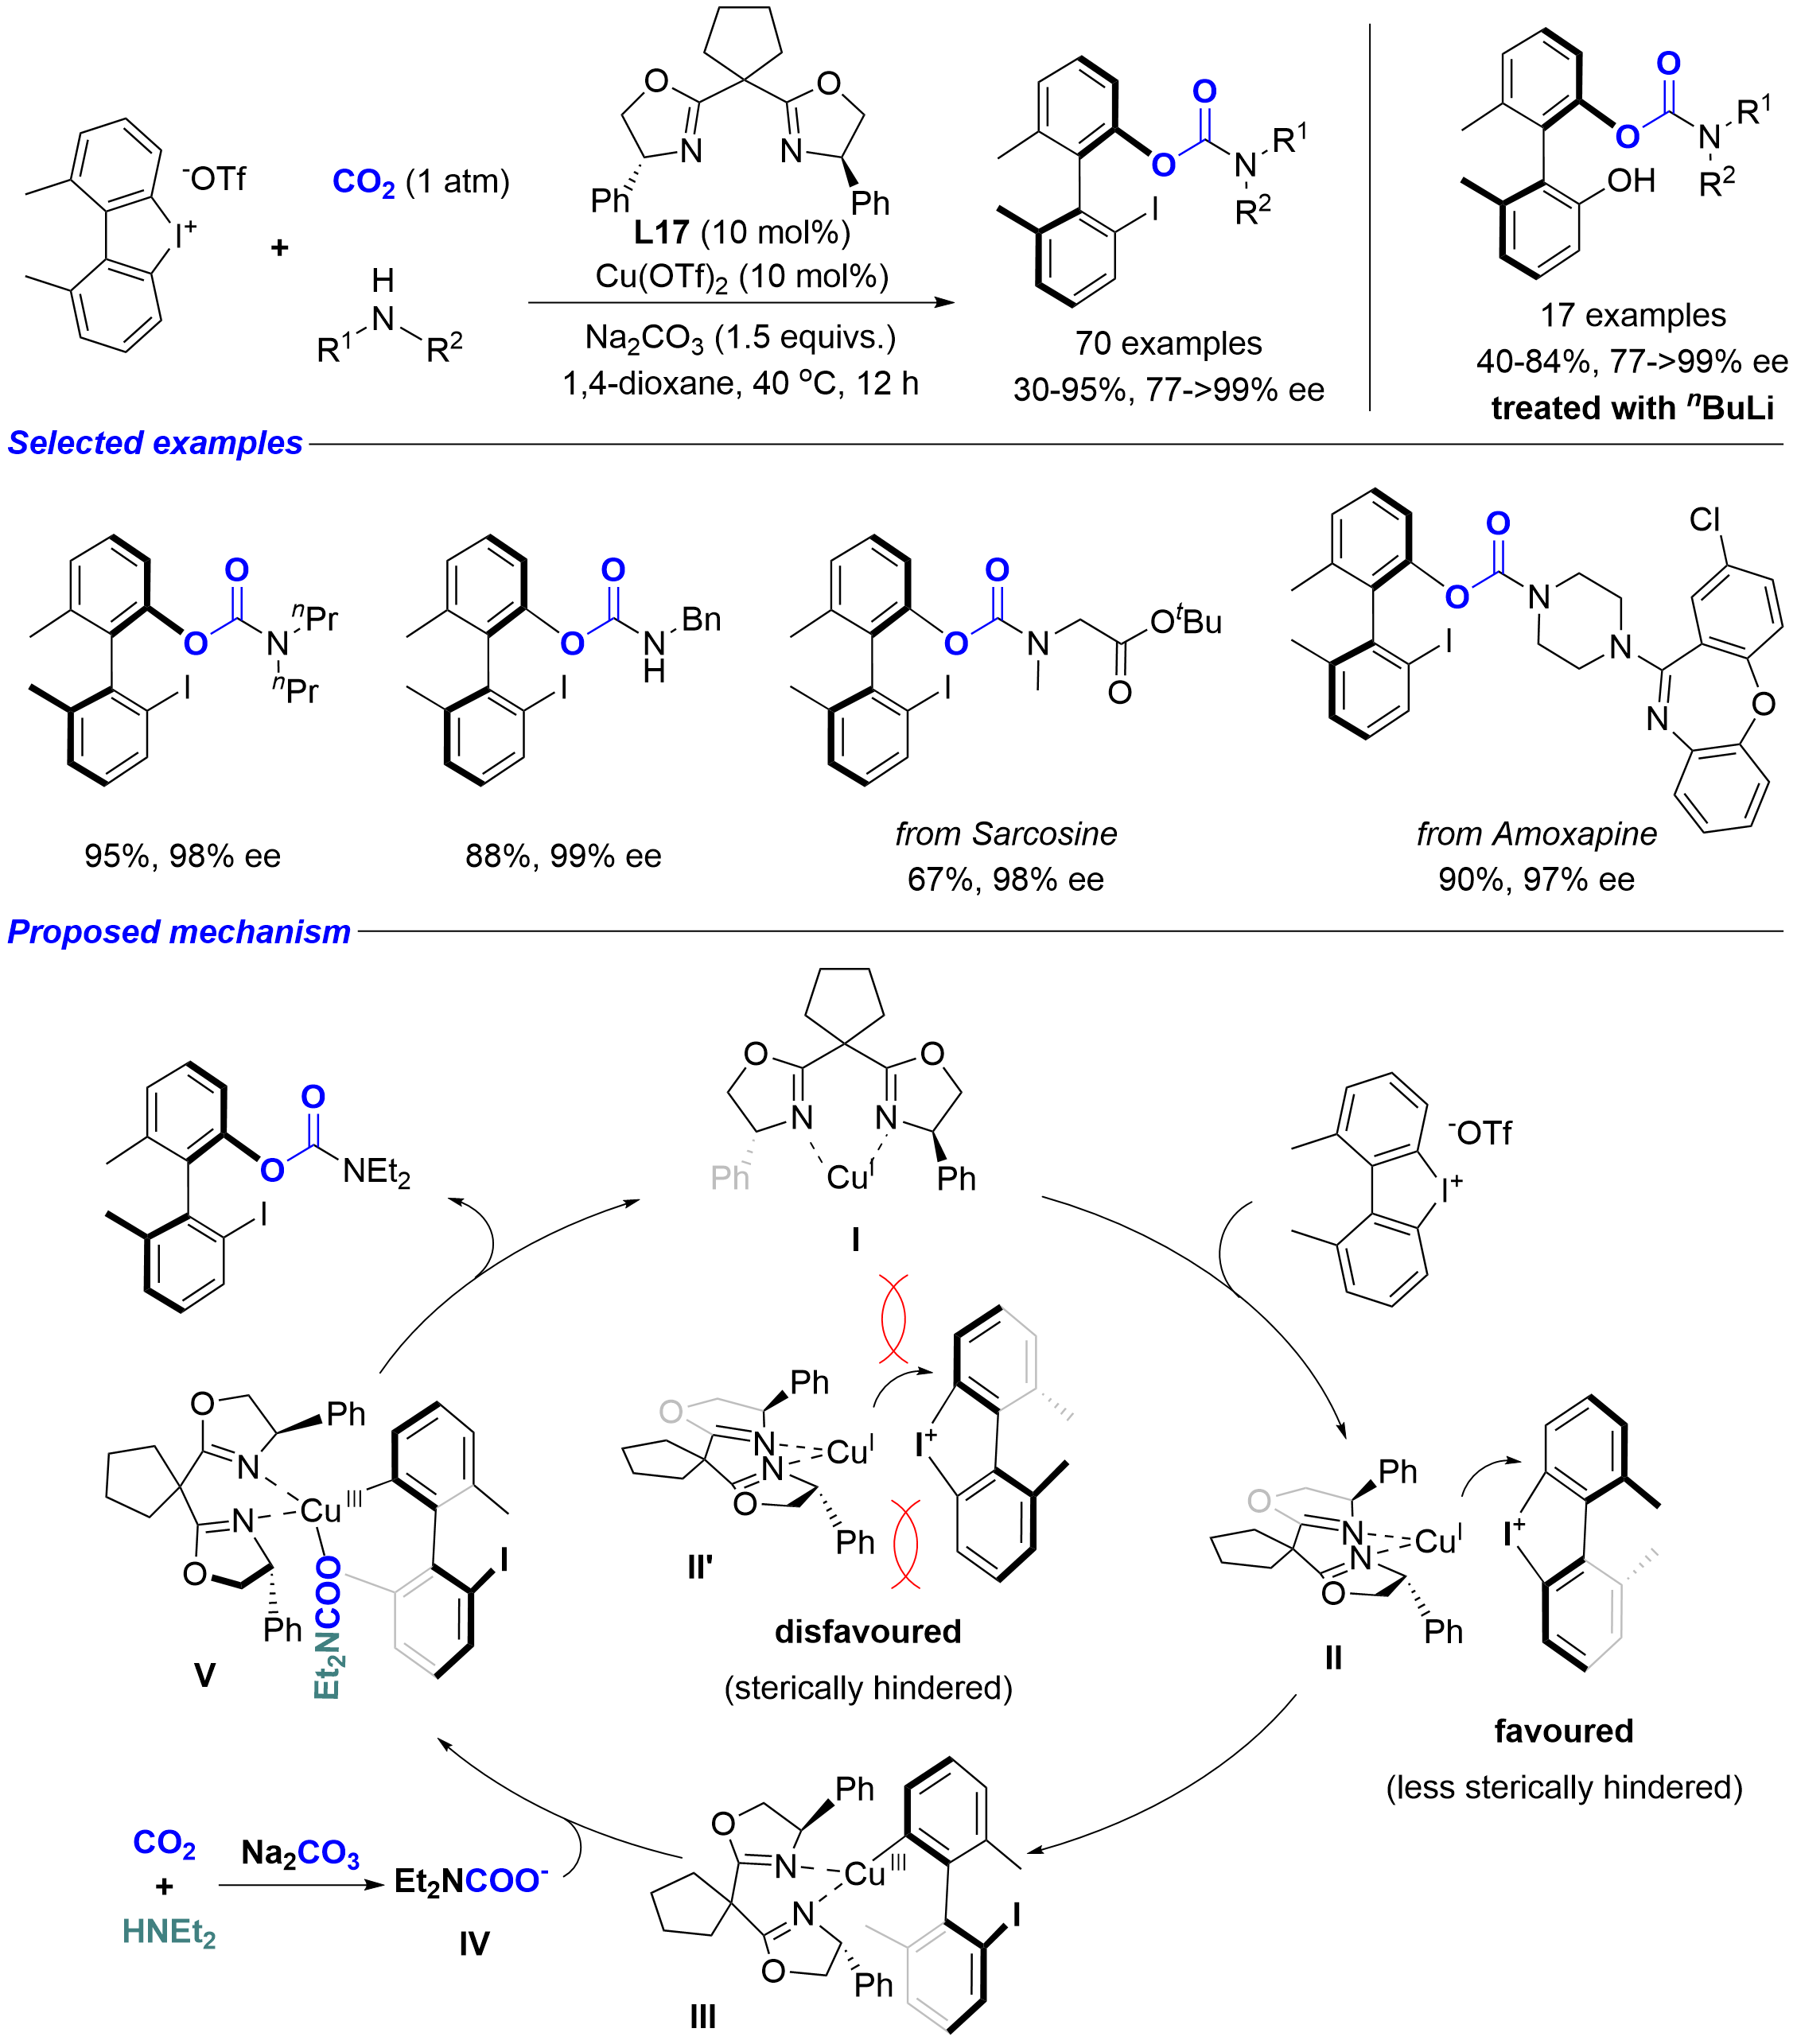
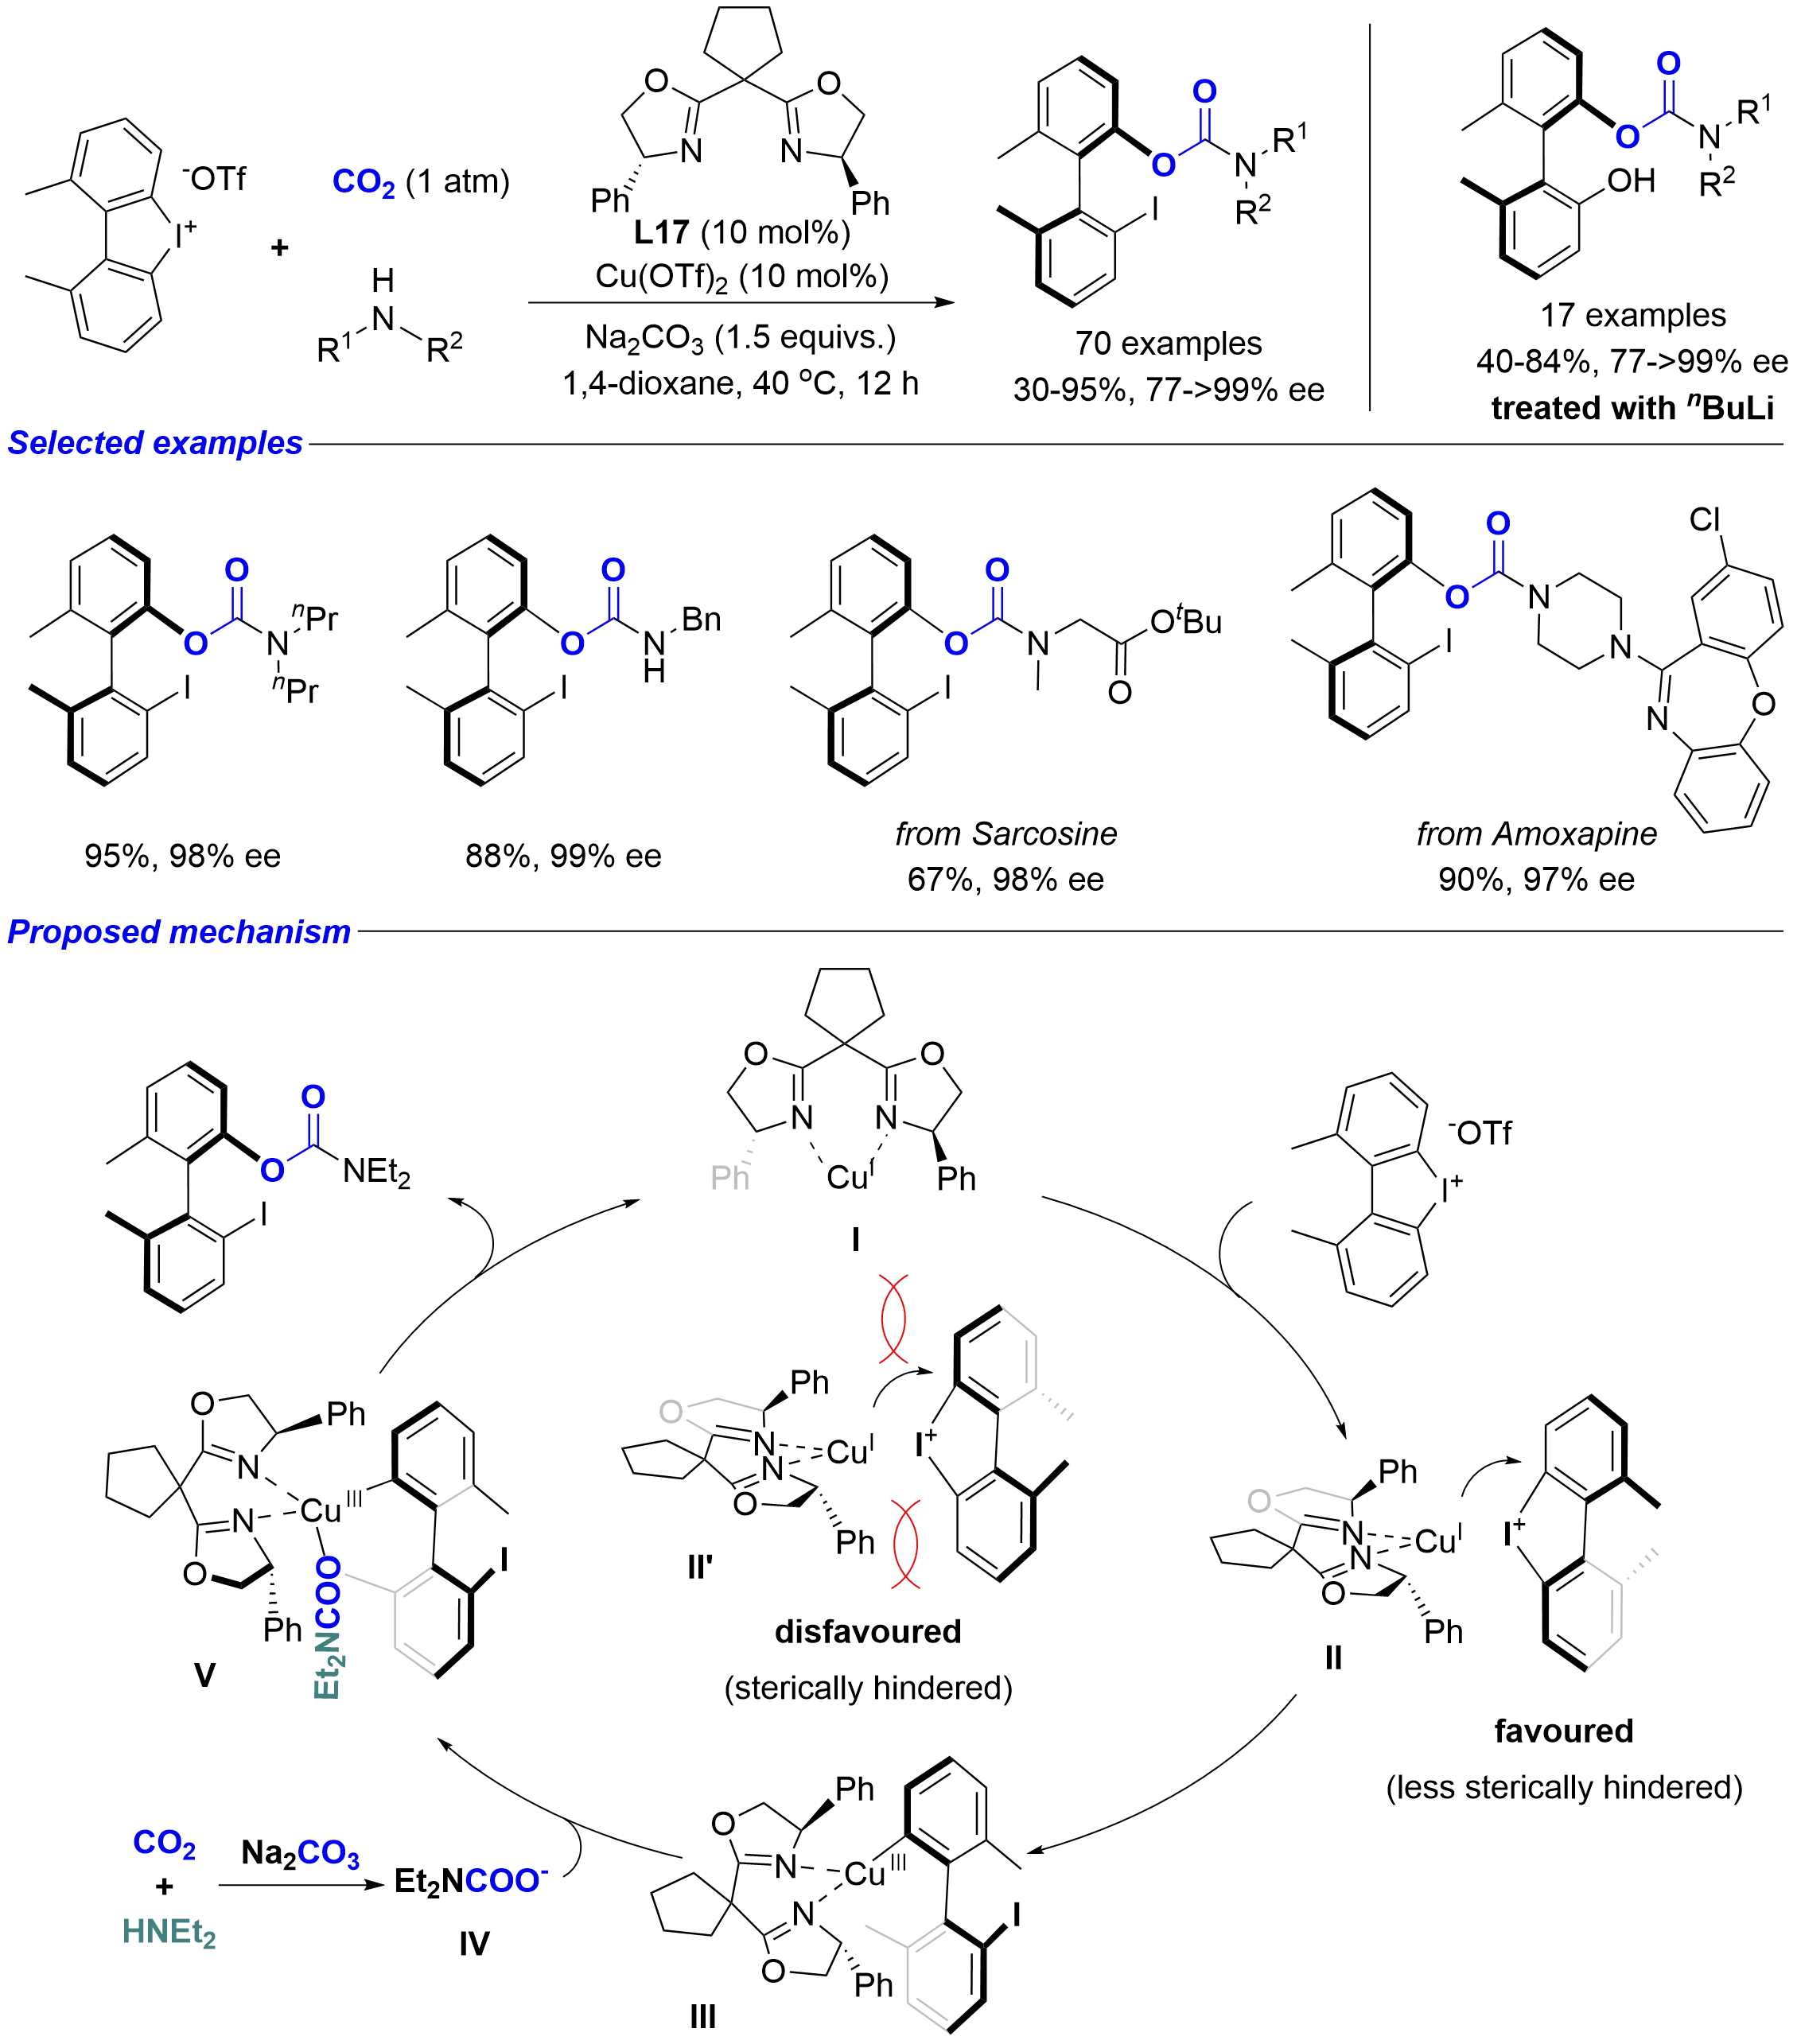
Scheme 19. Cu-catalyzed asymmetric three-component synthesis of axially chiral carbamates.
3. Asymmetric Synthesis of Chiral Heterocycles Involving CO2
3.1 Asymmetric synthesis of chiral 2-oxazolidinones with CO2
Chiral 2-oxazolidinones are important core scaffolds in pharmaceuticals and natural products. Their direct construction using CO2 as a C1 synthon offers advantages in green chemistry and atom economy. However, this transformation must simultaneously overcome the inherent inertness of CO2 and the challenges of controlling chemo- and enantioselectivity during cyclization. In recent years, multiple complementary catalytic strategies have been reported, significantly broadening the substrate scope.
As early as 2017, Zhou and co-workers reported an asymmetric A3 coupling-carboxylative cyclization reaction that directly constructs chiral 2-oxazolidinones from simple aldehydes, alkynes, and amines with CO2. However, this method was limited to 4-aryl-substituted frameworks[38]. To overcome this structural constraint, the same group reported a sequential catalytic asymmetric transfer hydrogenation-carboxylative cyclization reaction in 2021, which enabled the first highly enantioselective synthesis of 4-fluoroalkyl-substituted chiral 2-oxazolidinones (Scheme 20)[39]. This one-pot sequential strategy employed a chiral phosphoric acid-catalyzed asymmetric transfer hydrogenation to convert fluoroalkyl-substituted alkynyl ketimines into chiral propargylamine intermediates, followed by Ag2CO3- and DPG-cocatalyzed carboxylative cyclization in DMSO under 2.0 MPa CO2 at 25 °C, affording the desired products in up to 82% yield. The reaction exhibited good substrate scope, with trifluoromethyl-, difluoromethyl-, and pentafluoroethyl-substituted ketimines all participating smoothly to furnish the corresponding products. The resulting chiral oxazolidinones could be readily converted into β-fluoroalkyl-substituted amino alcohols.
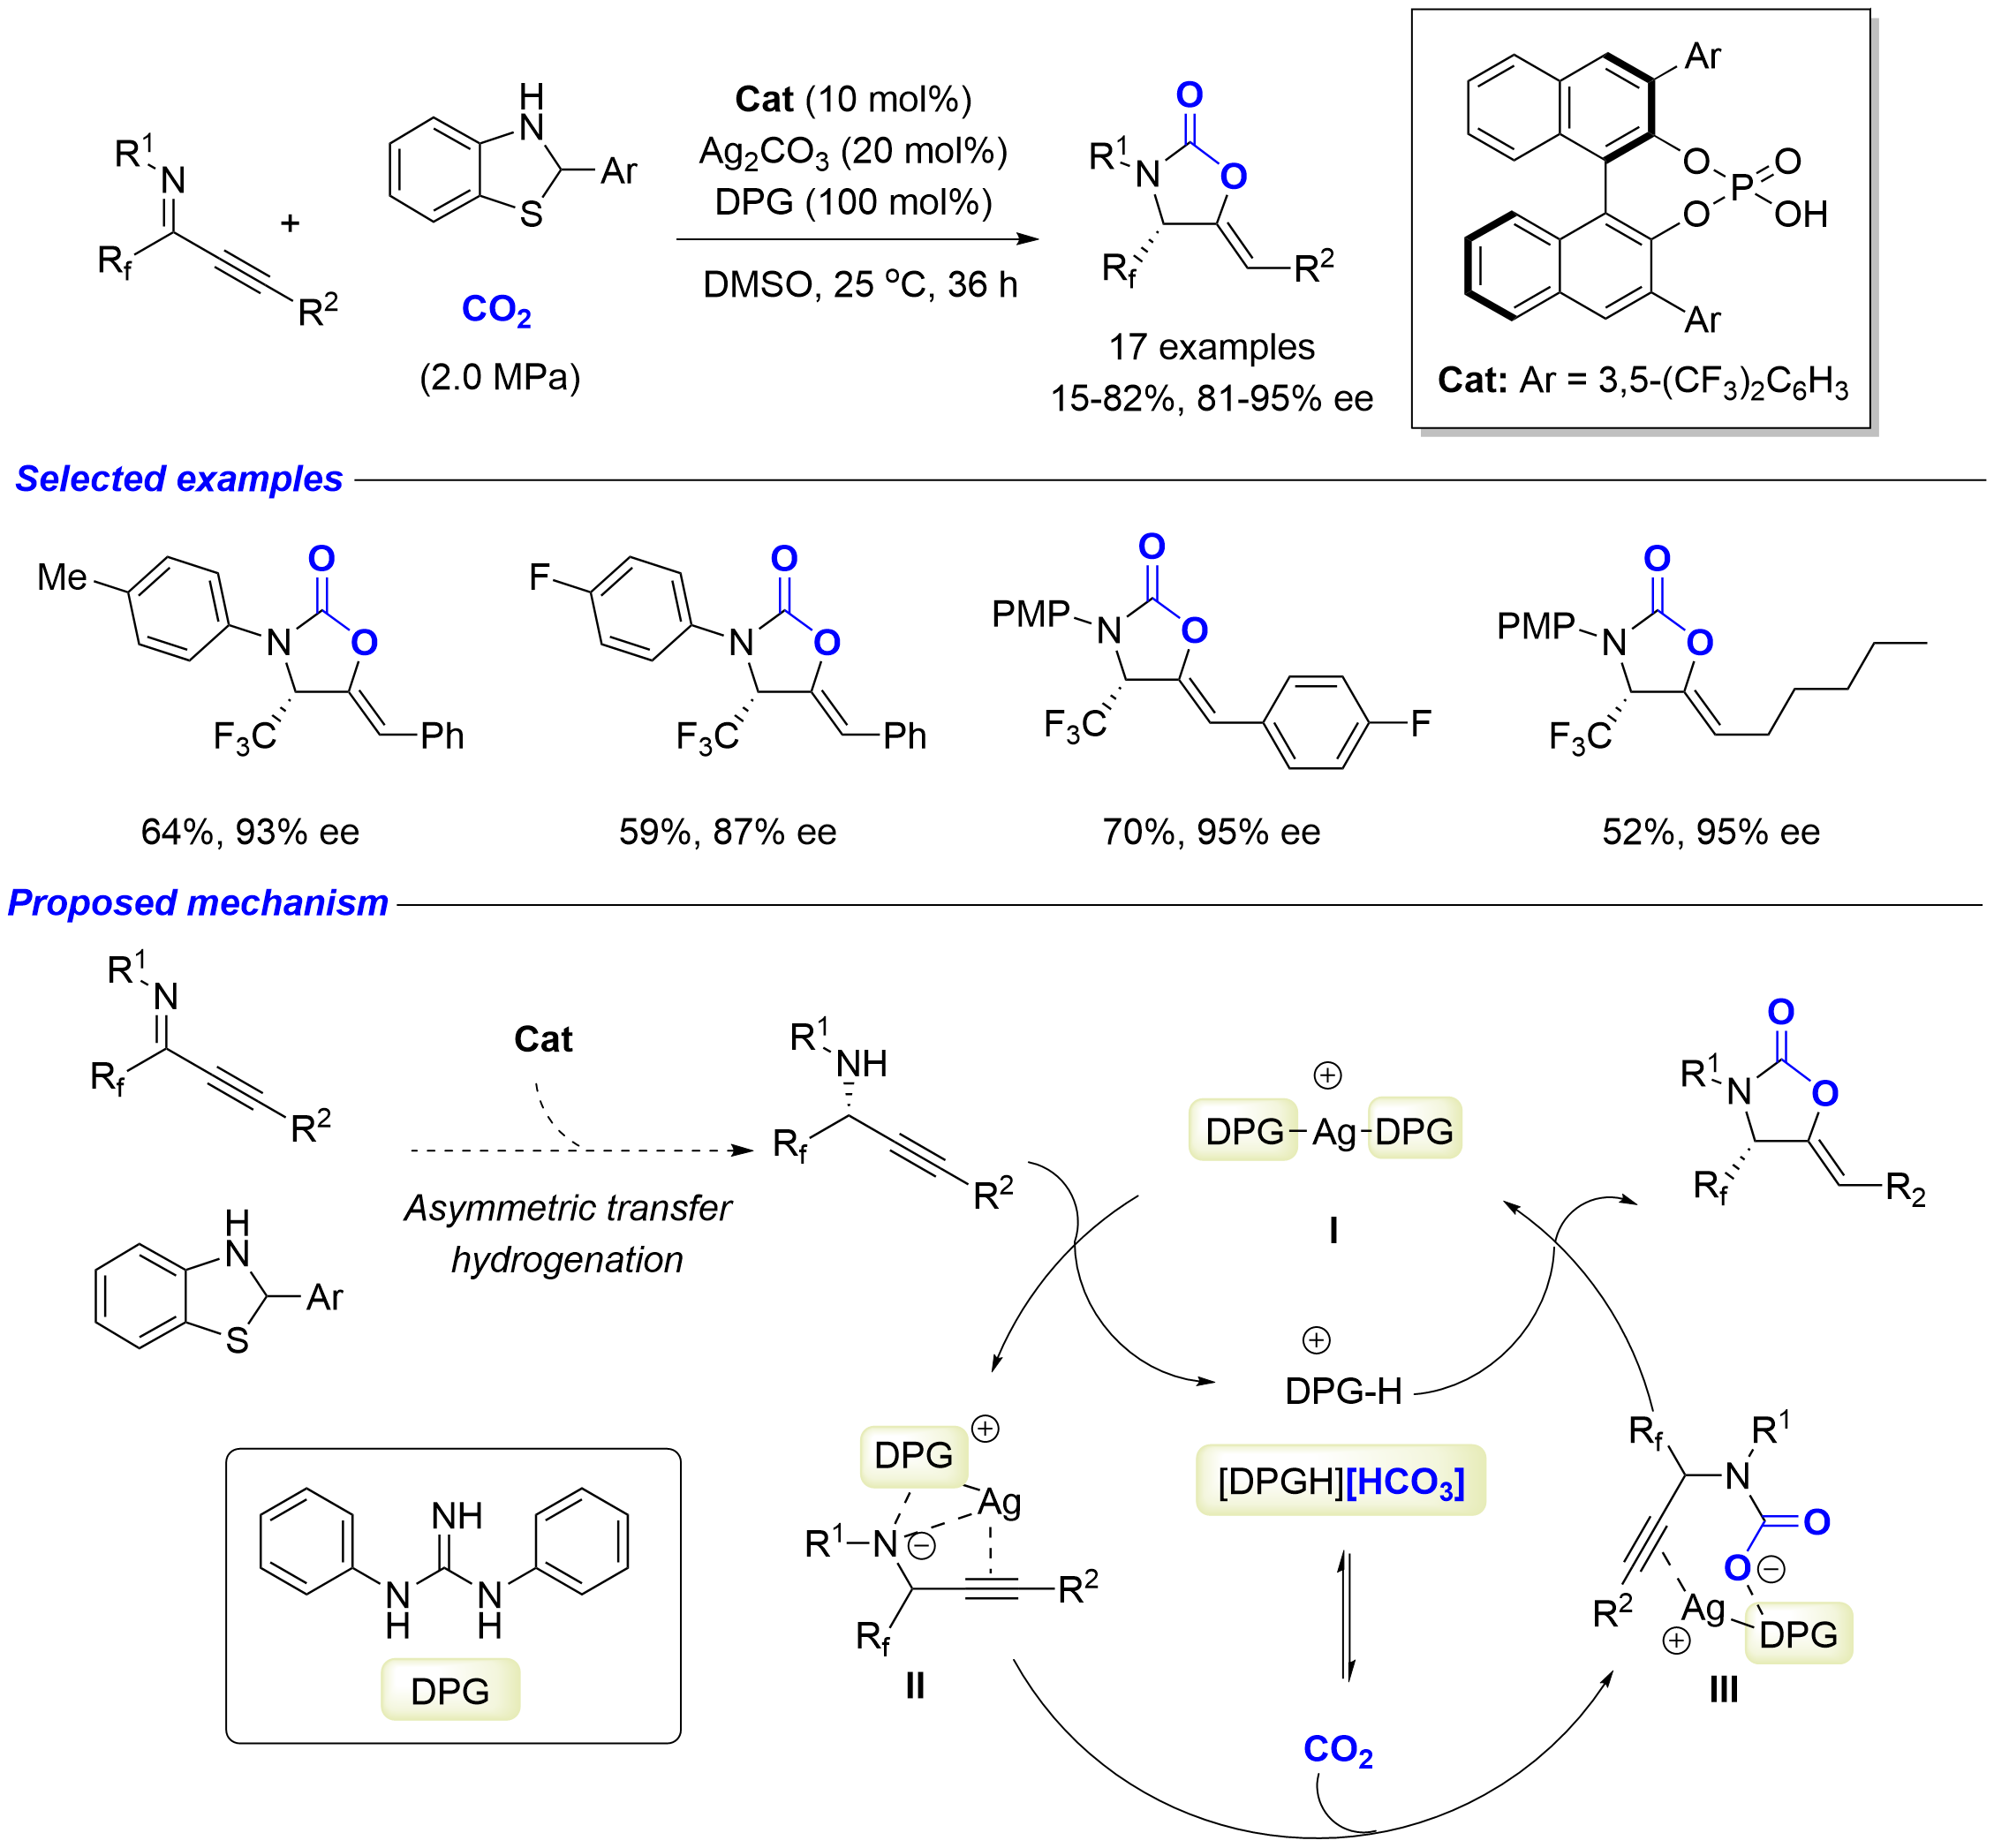
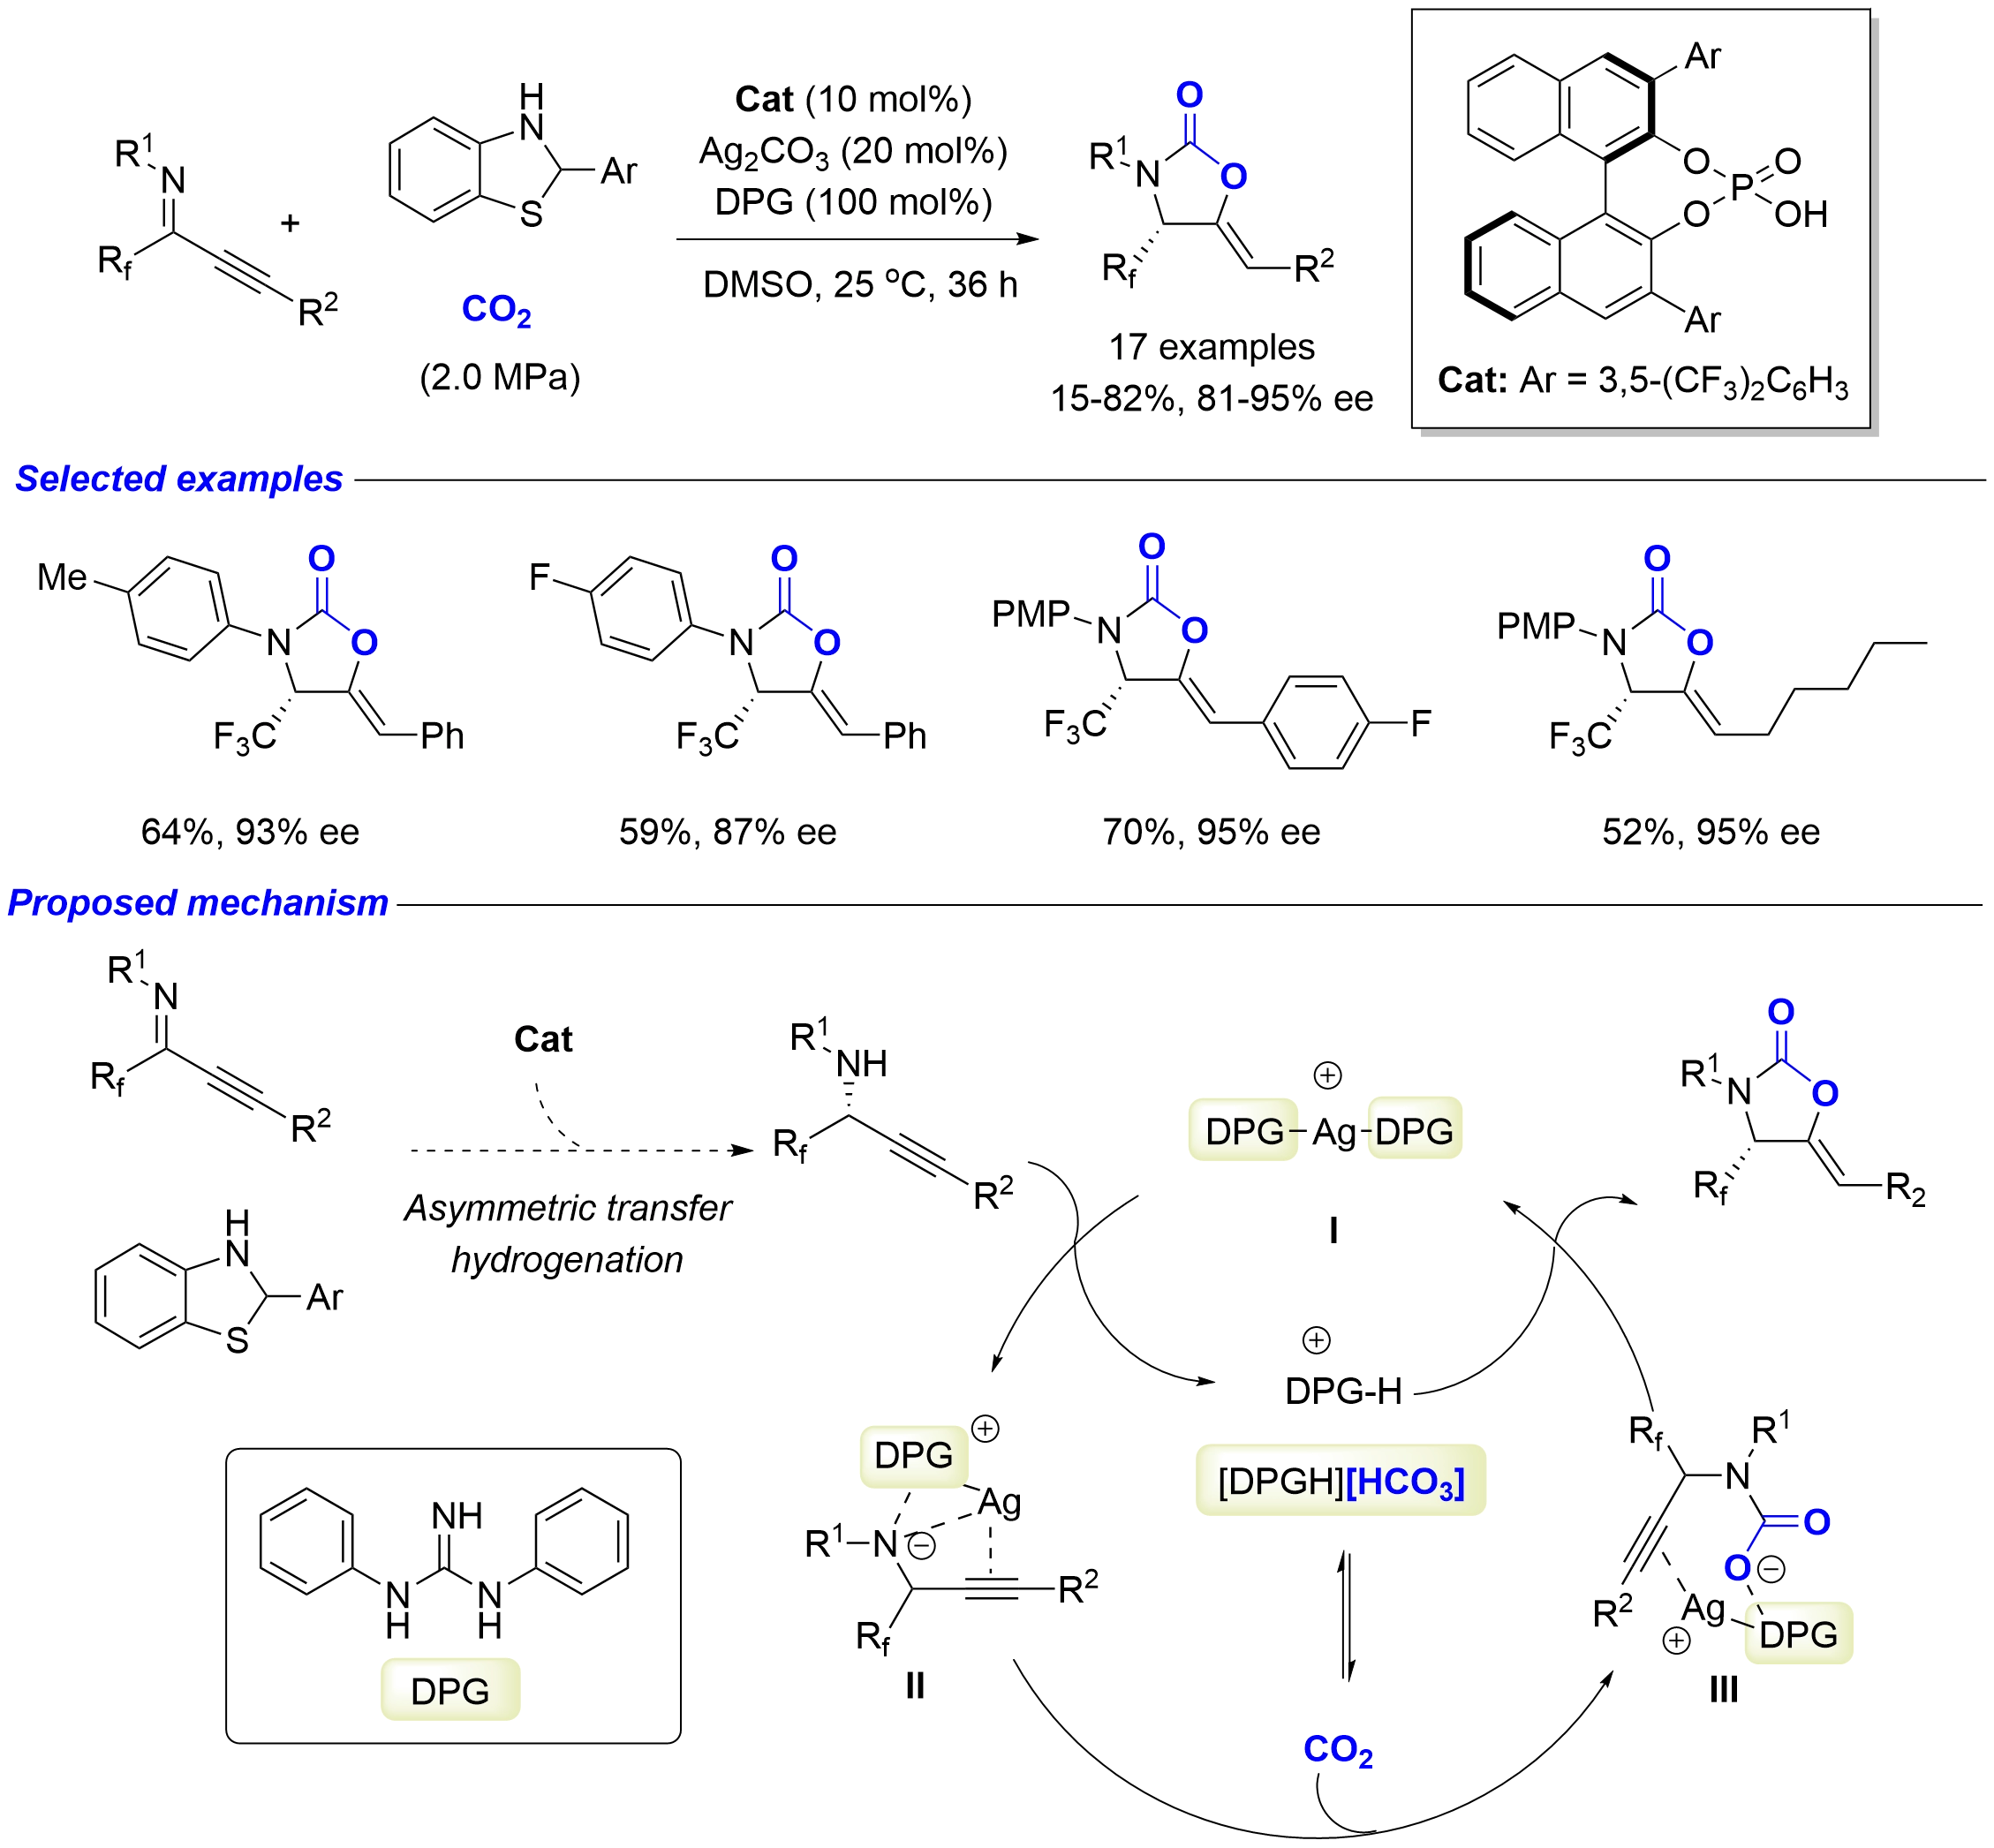
Scheme 20. One-pot asymmetric transfer hydrogenation/carboxylative cyclization to chiral 2-oxazolidinones.
Mechanistic studies revealed that the synergistic catalysis between 1,3-diphenylguanidine (DPG) and Ag(I) is crucial for the success of this carboxylative cyclization step[40]. DPG serves both as a base to facilitate the deprotonation of the propargylamine and as a ligand to stabilize the silver intermediate, thereby effectively overcoming the steric hindrance and electron-withdrawing effects imposed by the α-fluoroalkyl substituent. This enables efficient CO2 insertion to form a silver carbamate intermediate, which subsequently undergoes cyclization to furnish the 4-fluoroalkyl-2-oxazolidinone.
Notably, the above-mentioned studies mainly yielded chiral oxazolidinones bearing a 5-arylidene moiety, and were unable to prepare chiral 2-oxazolidinones with an exocyclic methylene motif. These compounds are valuable platform molecules that can be converted into structurally diverse 2-oxazolidinones via versatile diversifying reactions of the terminal alkene moiety. In 2024, He and coworkers developed a copper-catalyzed asymmetric propargylic substitution reaction leveraging CO2 shuttling and fixation technology (Scheme 21)[41].
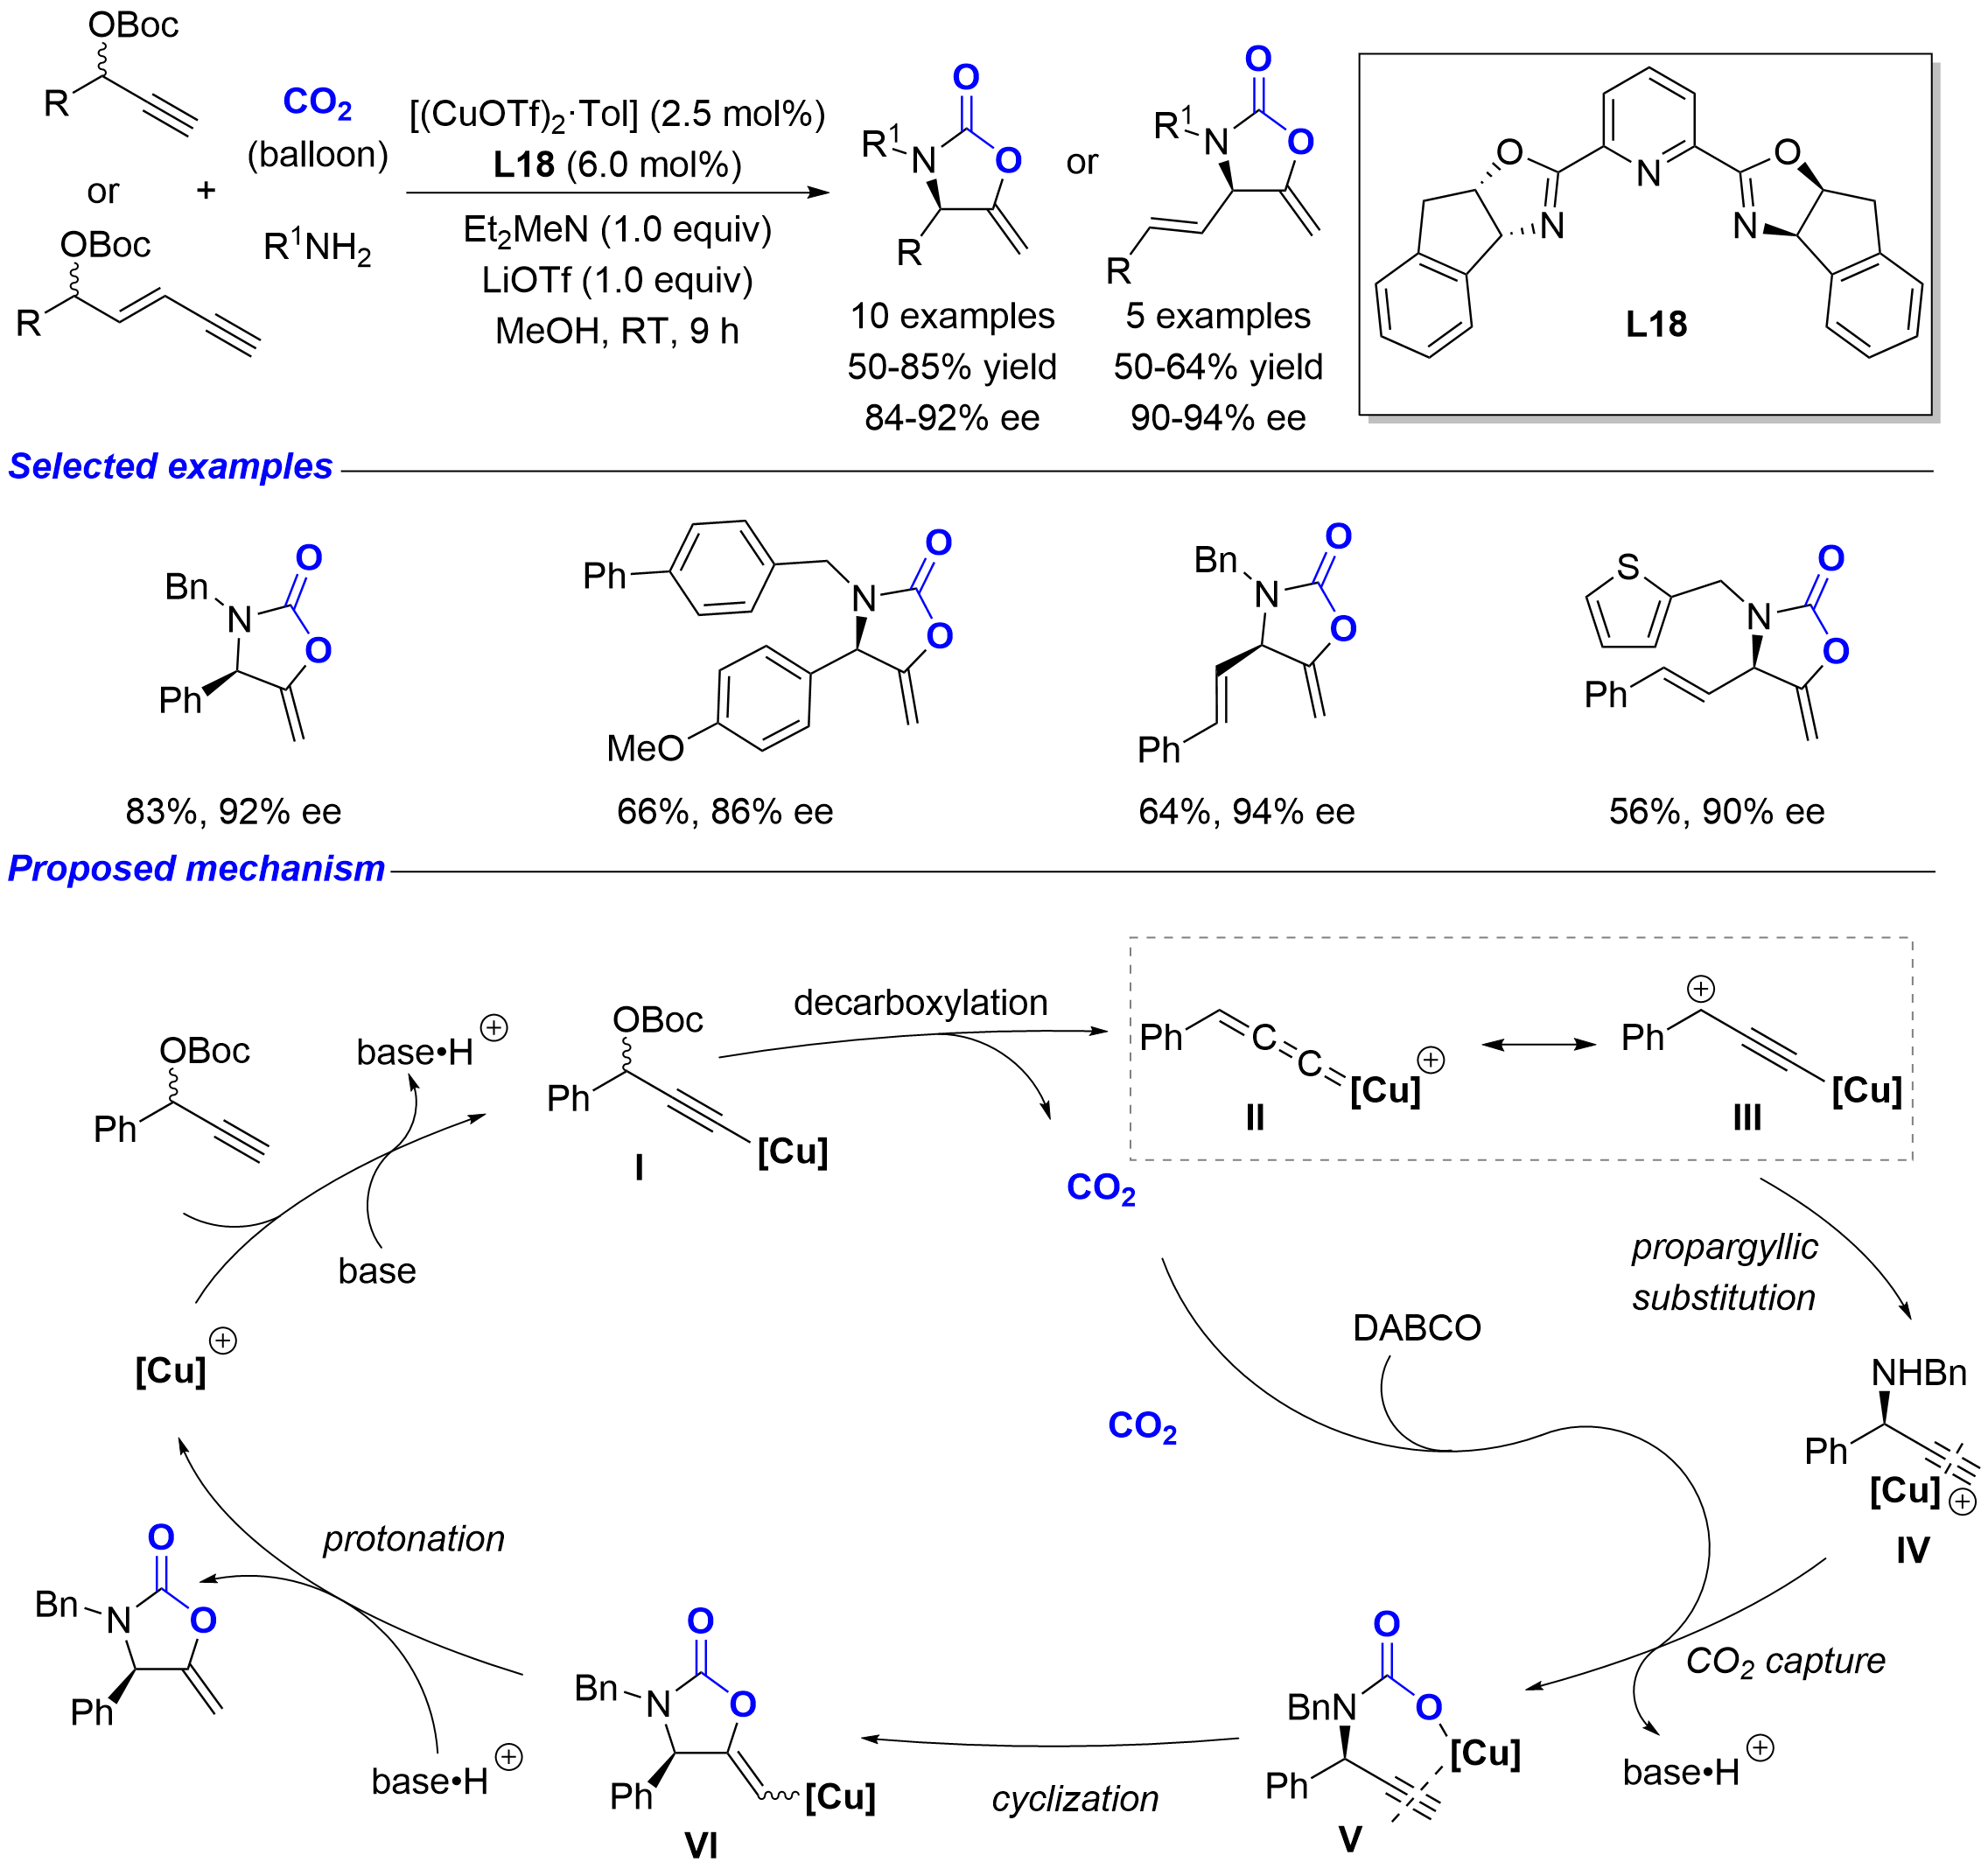
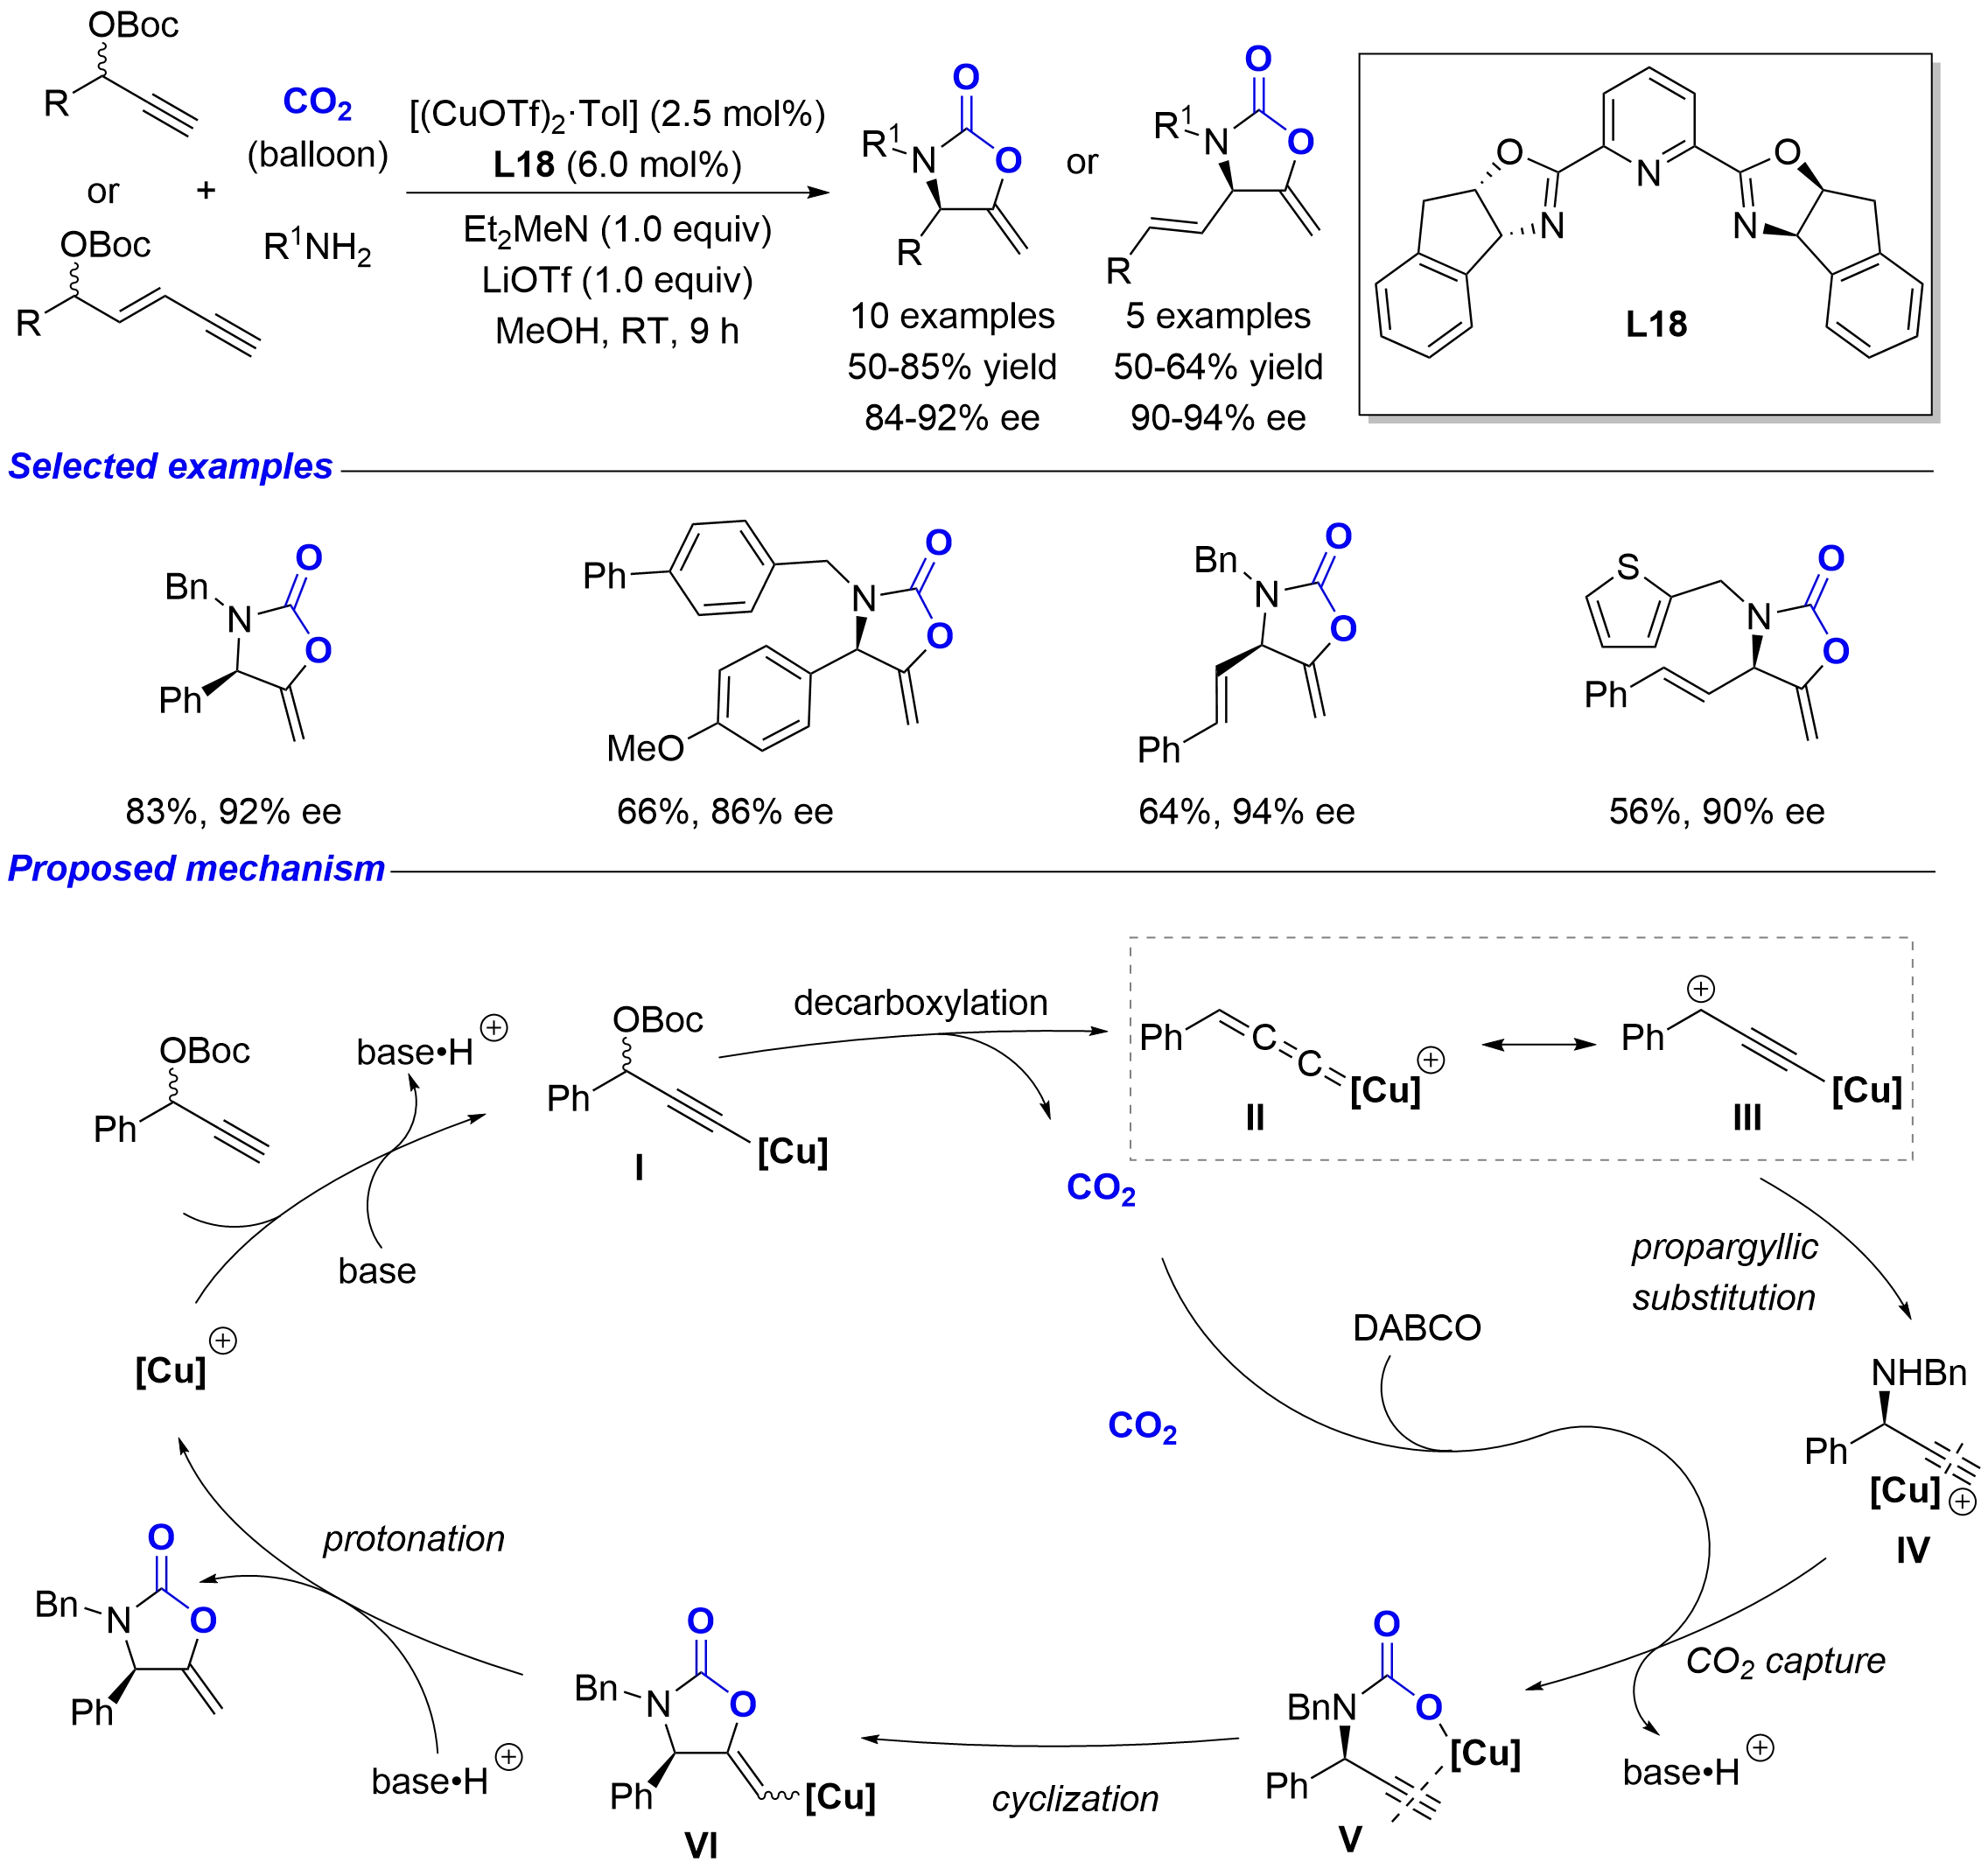
Scheme 21. Cu-catalyzed CO2-shuttling asymmetric propargylic substitution.
Using conjugated enyne esters or propargylic carbonates as electrophilic substrates and primary amines as nucleophiles, in the presence of a chiral PYBOX ligand and a copper catalyst, the CO2 shuttling process was efficiently promoted, affording chiral oxazolidinone products in 50-85% yield with up to 94% ee. By replacing DABCO with Et2MeN, a three-component asymmetric propargylic substitution under atmospheric CO2 was realized with comparable results. Substituting 12CO2 with 13CO2 enabled the convenient synthesis of 13C-labeled oxazolidinones, providing a practical tool for isotope tracing studies. Mechanistic studies revealed that the chiral PyBox/Cu(I) catalyst coordinates with propargylic carbonate, and cleavage of the carbonate leaving group gives electrophilic Cu-allenylidene species II. Then, enantioselective propargylic amination gives intermediate IV, which sequentially captures CO2 to form V. The intramolecular carboxylative cyclization of V via nucleophilic attack of the oxygen anion on the alkene (or alkyne) moiety gives oxazolidinone intermediate VI, which releases the chiral product and regenerates the Cu(I) catalyst. In the CO2 shuttling pathway, DABCO acts as both a base for propargylic substitution and a capture agent for the released CO2.
In the same year, Wang, Cai and coworkers developed a dinuclear copper-catalyzed asymmetric propargylic amination-carboxylative cyclization tandem reaction (Scheme 22)[42]. Using propargylic carbonates and alkylamine hydrochlorides as substrates, a chiral dinuclear copper complex bearing a benzo[c]quinoline bisoxazoline framework as the catalyst, tBuONa as the base, and Ag2CO3 as the additive, the reaction proceeded in CF3CH2OH at room temperature under atmospheric CO2 for 36 h to afford a series of chiral 2-oxazolidinones bearing exocyclic methylene groups in one pot, with yields of up to 88% and enantioselectivities of up to 96% ee. This work extends the application of dinuclear copper complexes in asymmetric catalysis and offers a strategy for the construction of chiral heterocycles using CO2 as a C1 building block.
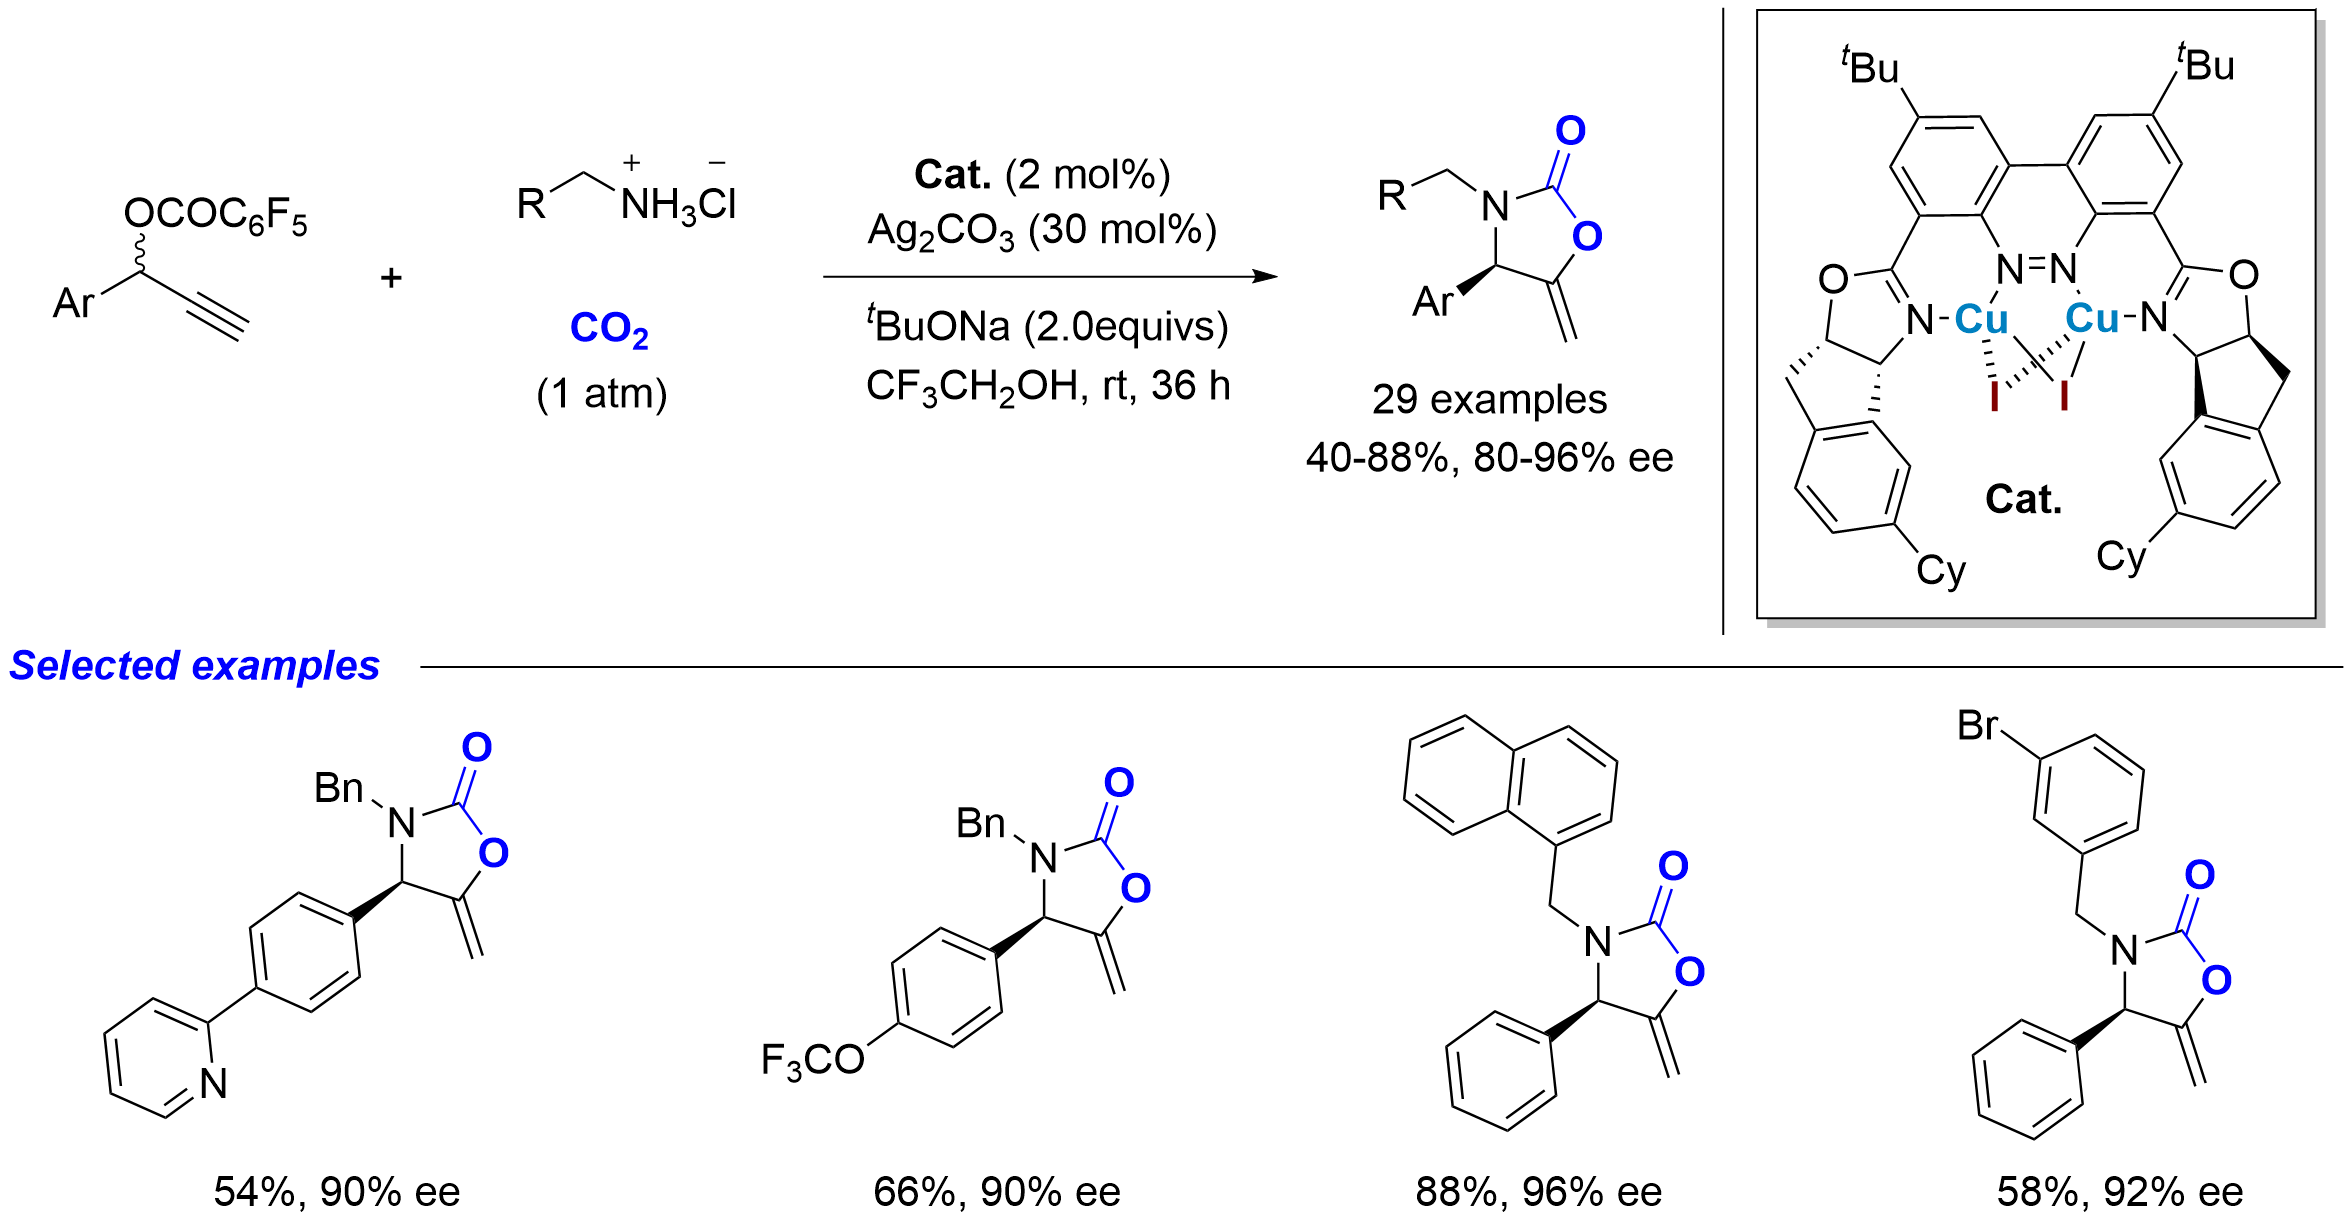
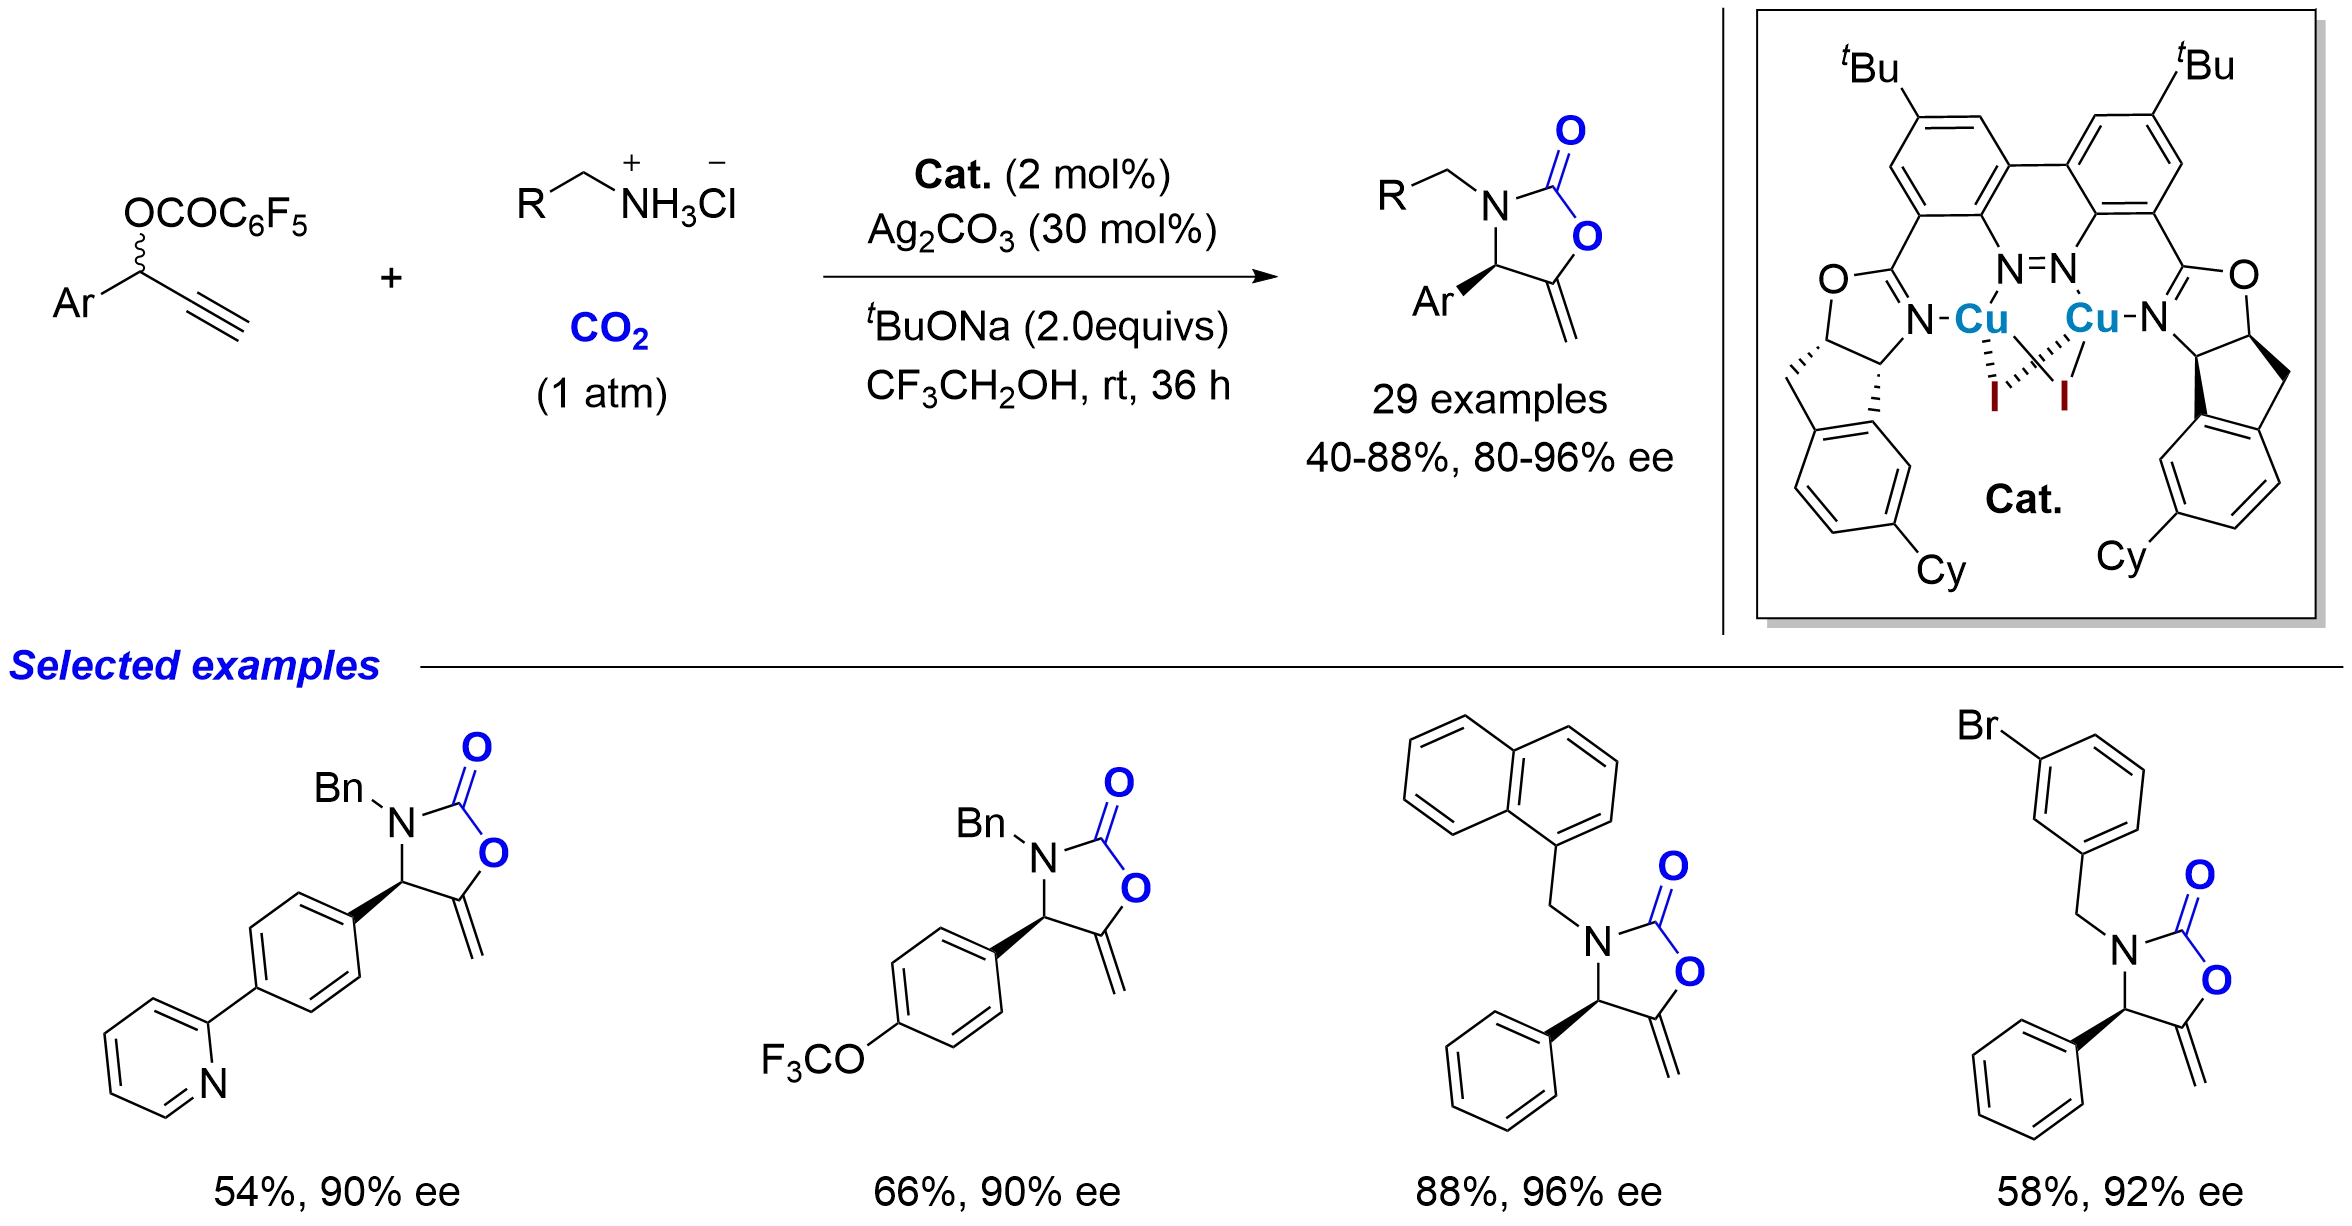
Scheme 22. Dinuclear copper-catalyzed asymmetric tandem propargylic amination/carboxylative cyclization.
Despite the aforementioned advances, the construction of chiral 5-methylidene-2-oxazolidinones bearing an α-aliphatic substituent, or even those with α-tetrasubstituted stereocenters, remains a challenging task, due to the inherent difficulty of asymmetric Cu-catalyzed propargylic amination of α-aliphatic secondary and α-tertiary propargylic electrophiles. In 2024, Zhou and co-workers developed a new type of sterically confined PYBOX ligands bearing C4 substituents to achieve unprecedented, highly enantioselective asymmetric Cu-catalyzed propargylic amination reactions of simple aryl- or aliphatic ketone-derived propargylic carbonates, enabling the construction of a wide range of α-tertiary ethynylamines with high efficiency[43]. Building on these results, they further explored a tandem asymmetric propargylic amination-carboxylative cyclization reaction of propargylic esters with primary amines, thereby expanding the scope of diversity-oriented synthesis of chiral 2-oxazolidinones (Scheme 23)[44].
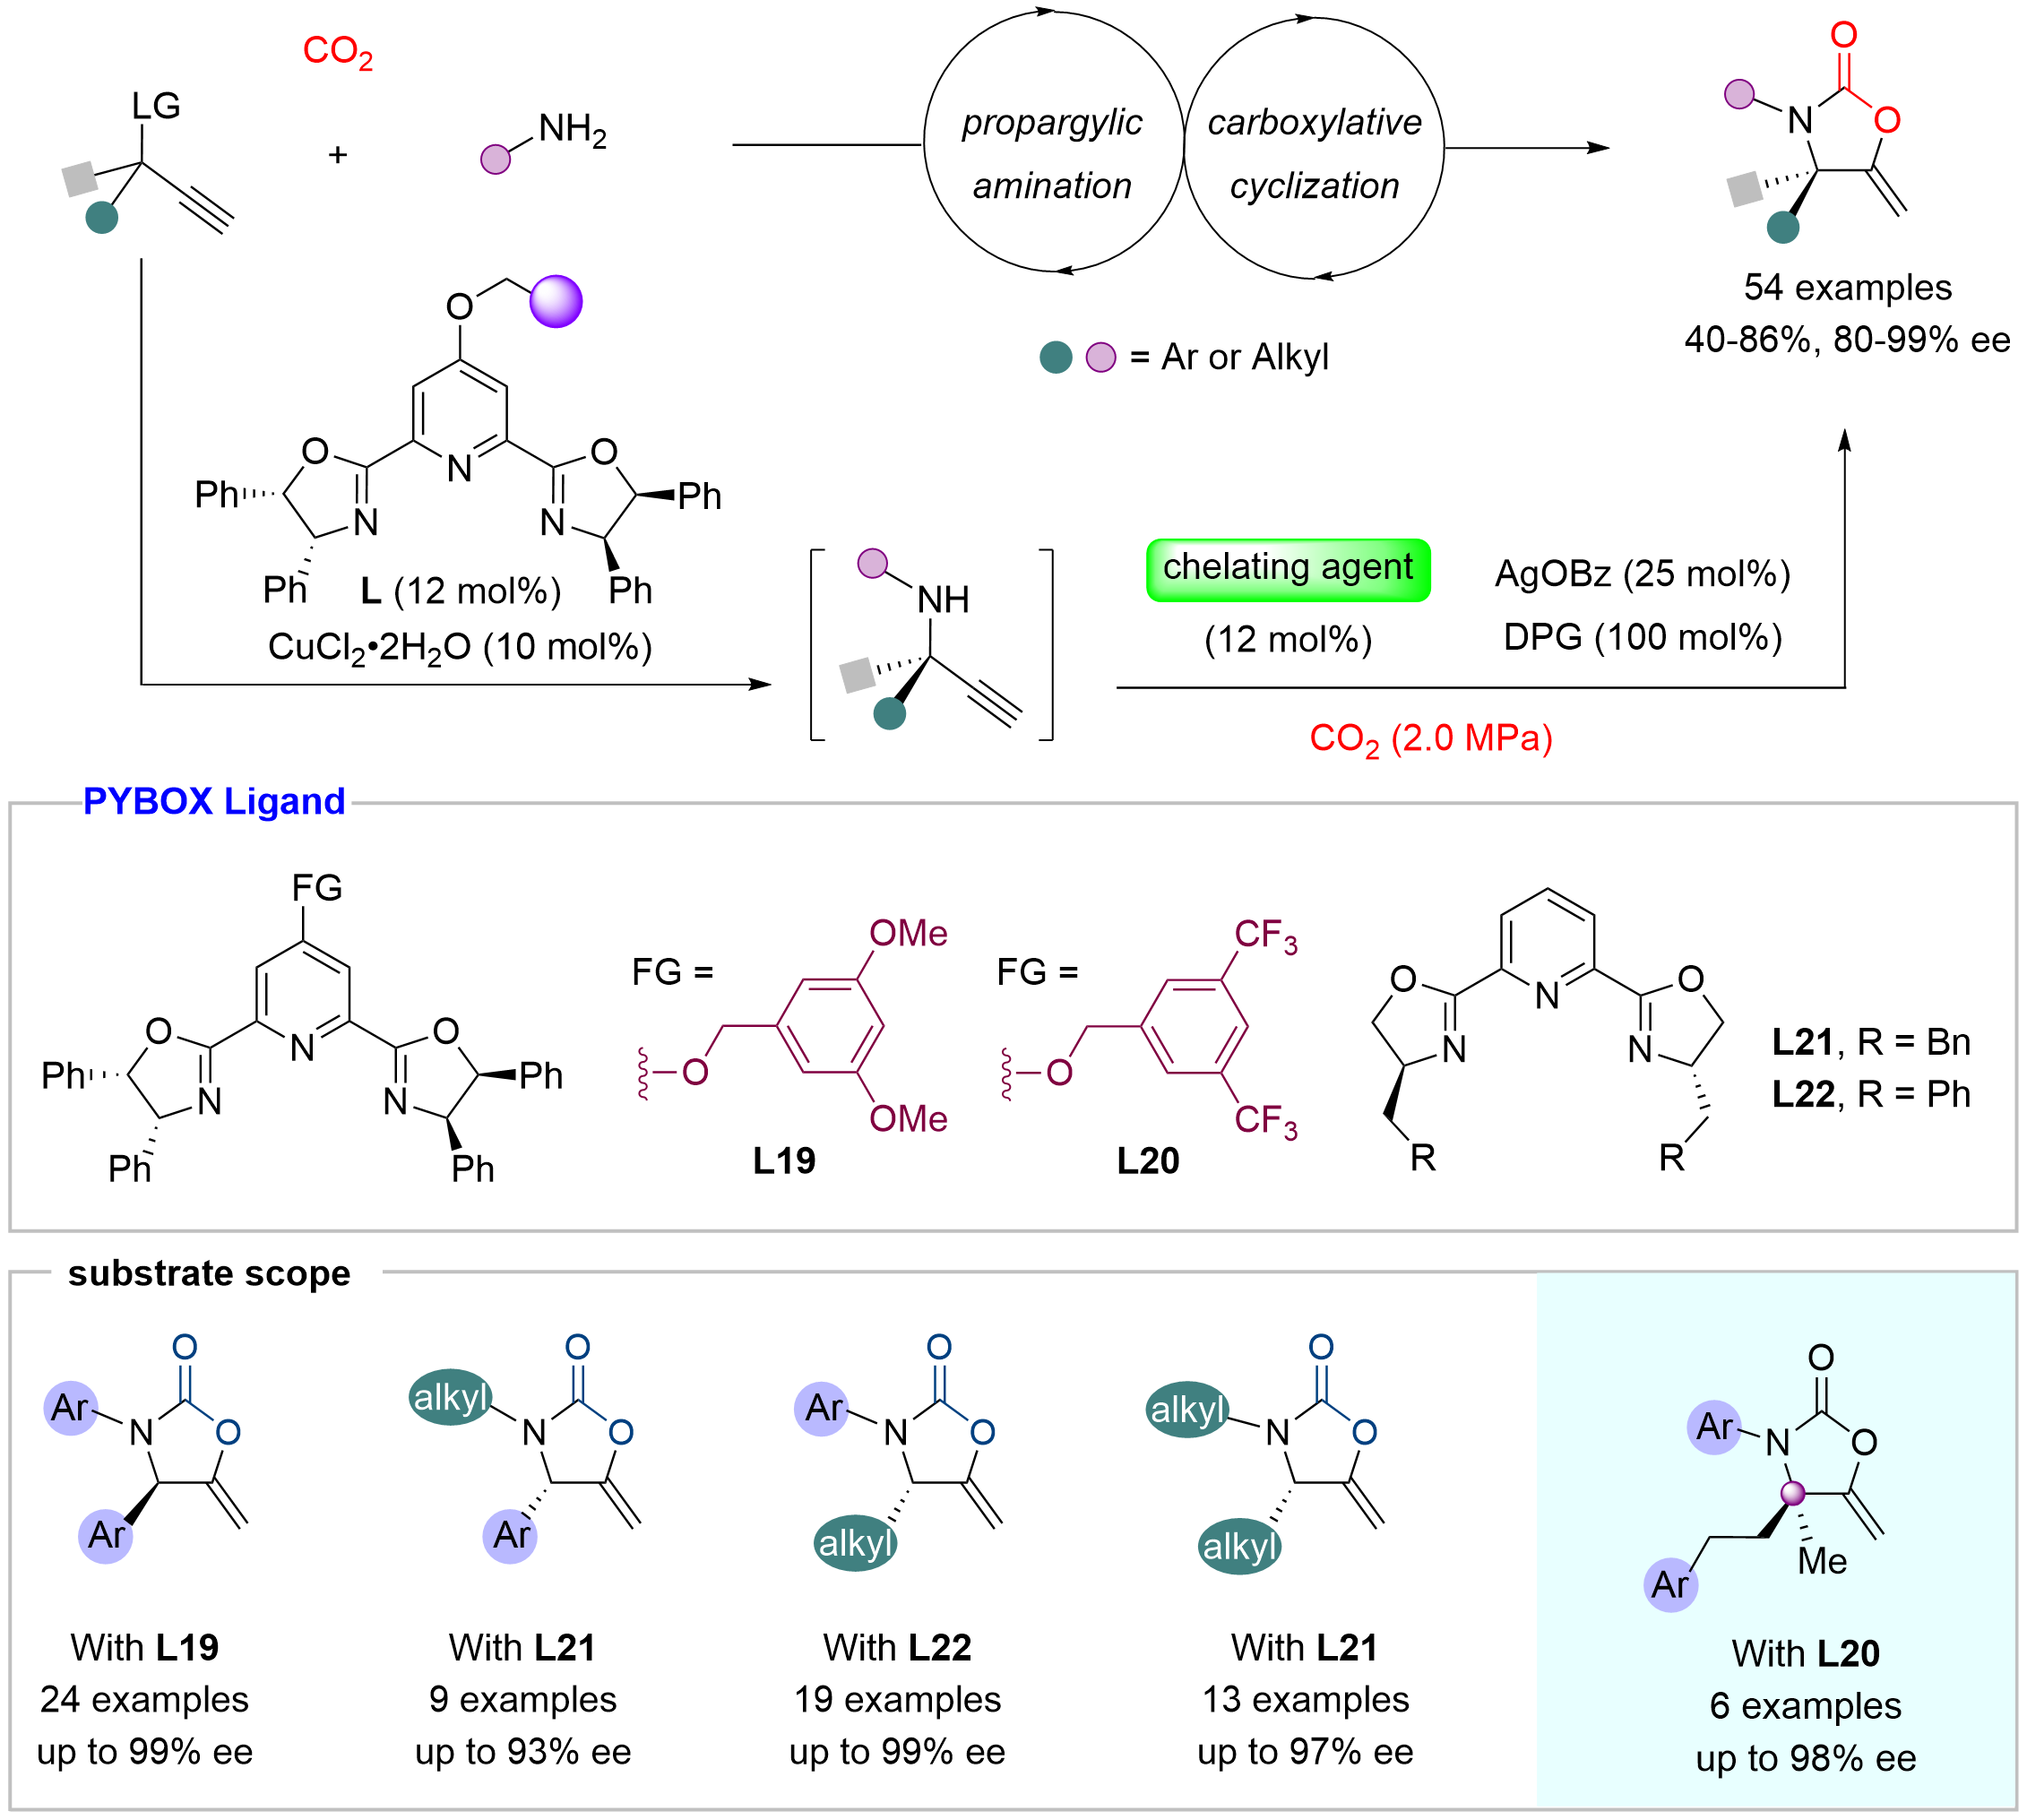
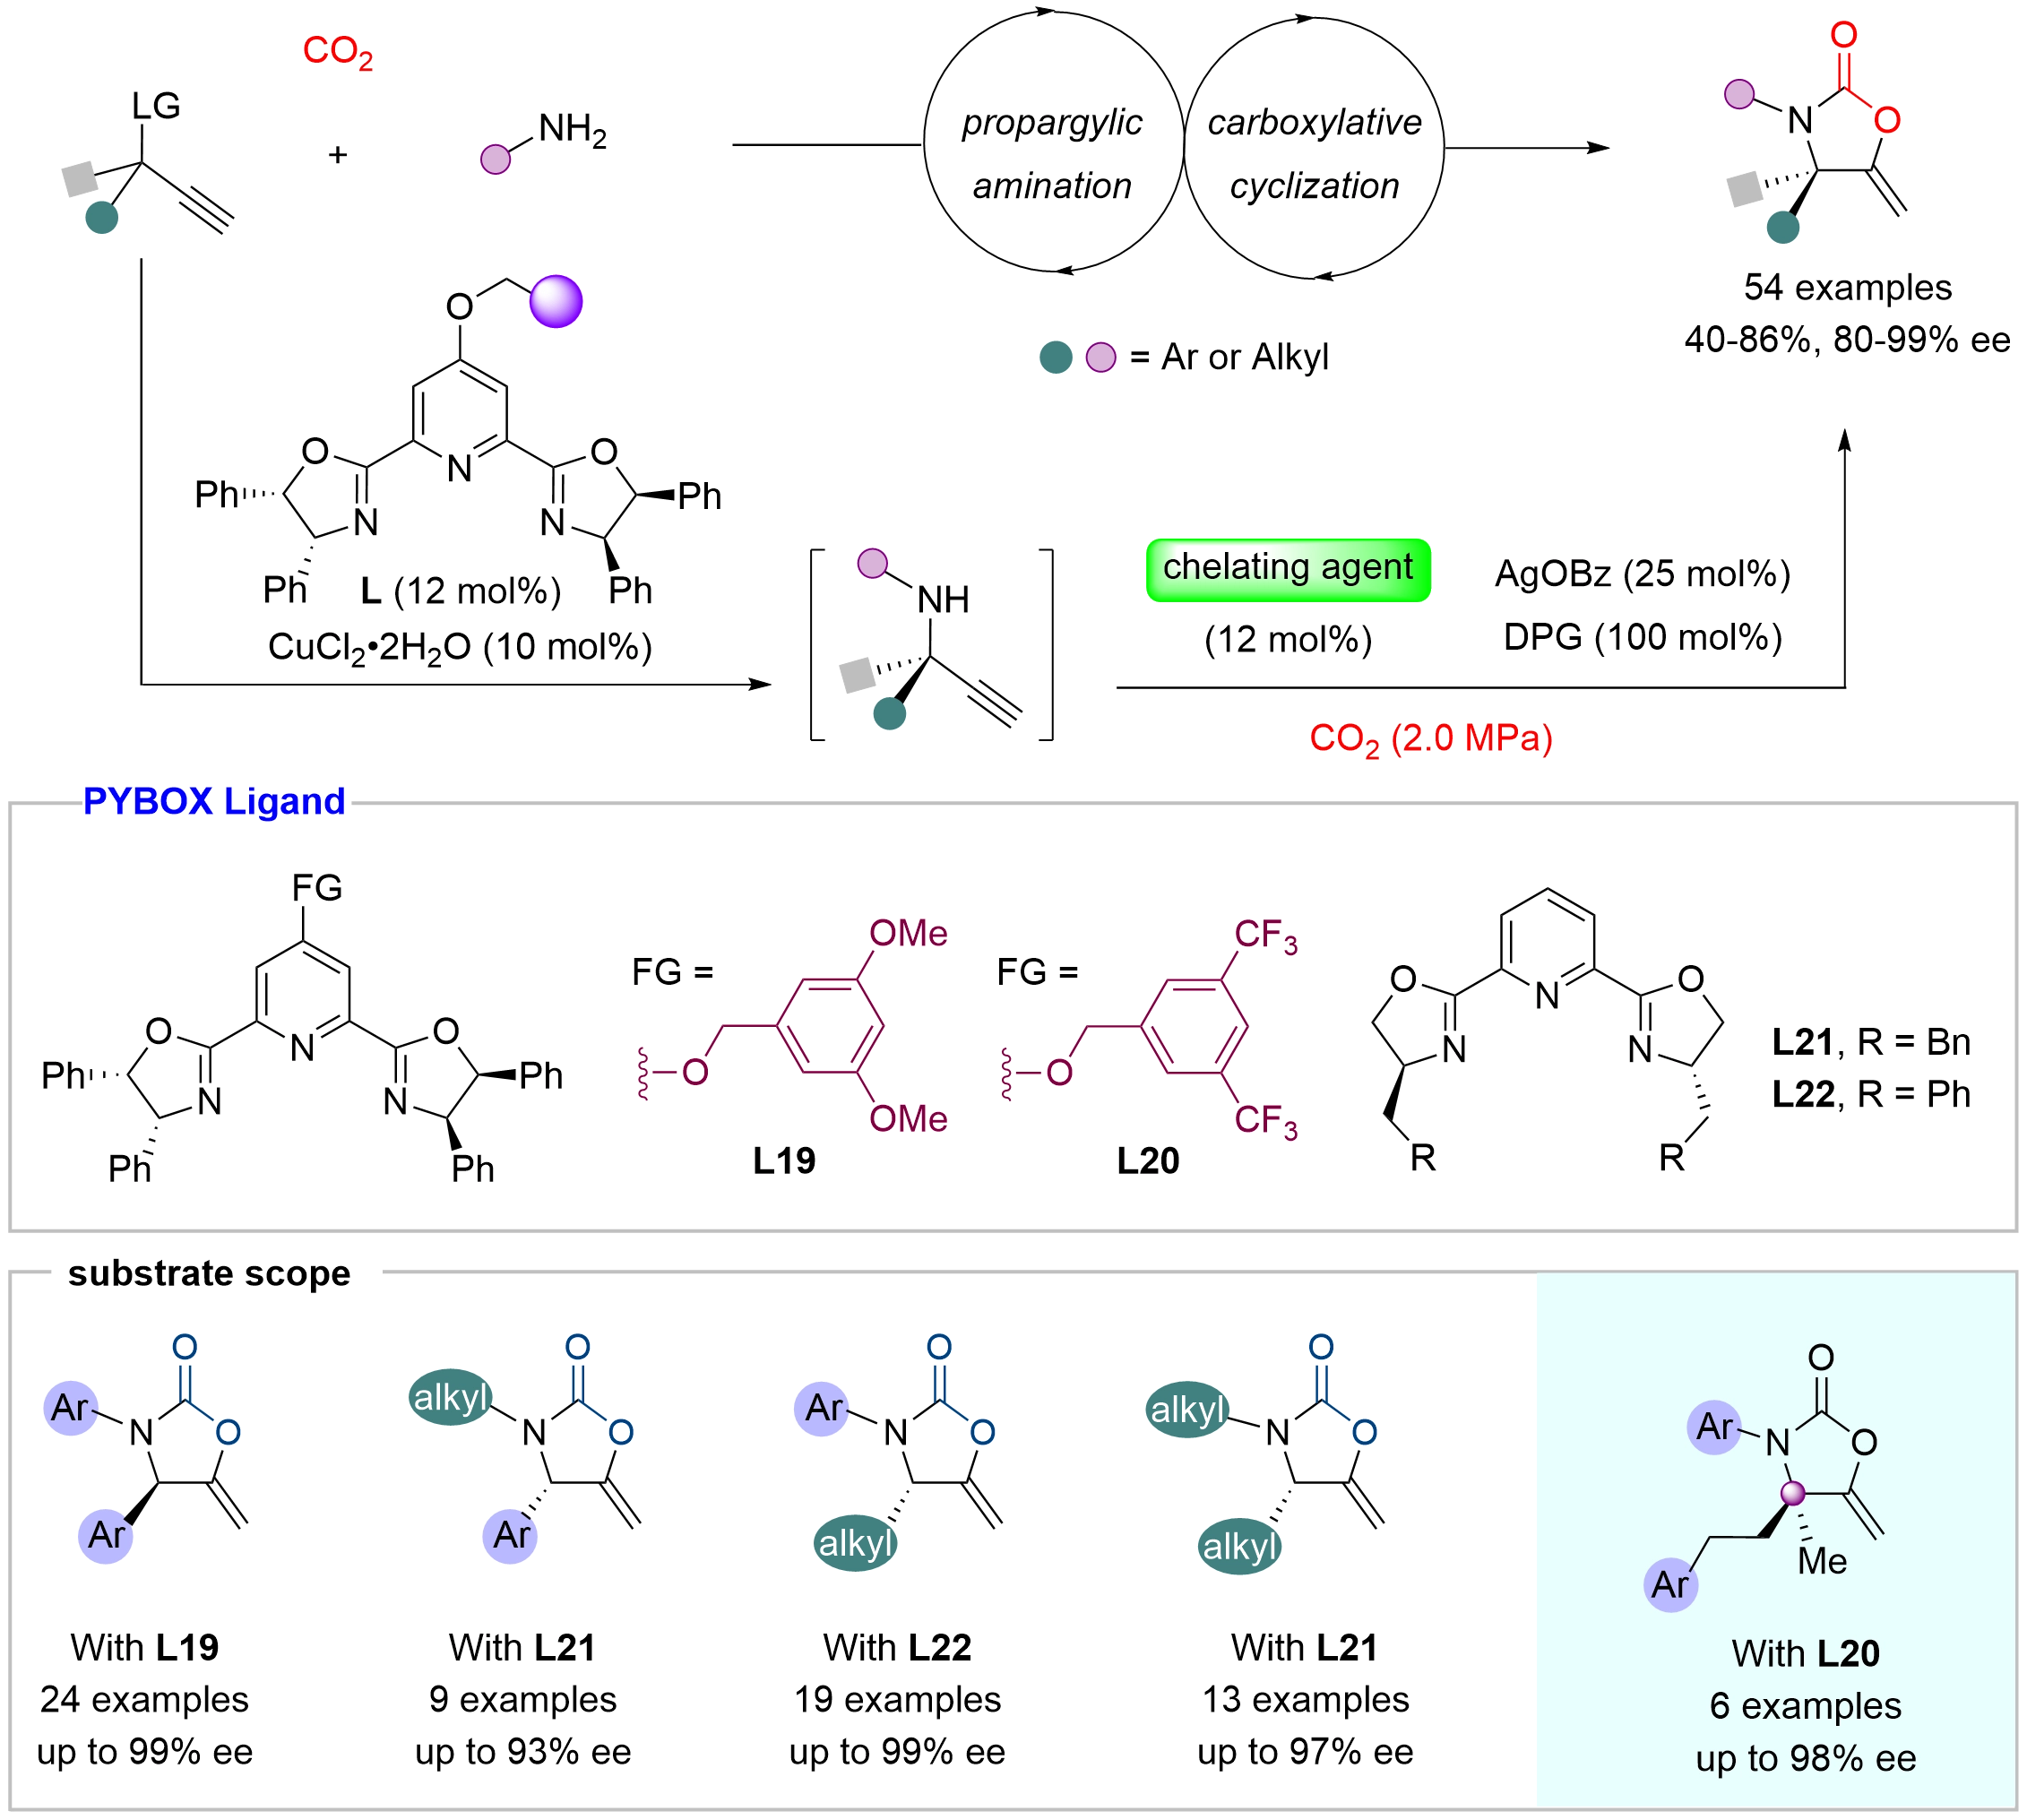
Scheme 23. Cu/Ag-tandem asymmetric propargylic amination/carboxylative cyclization.
A new type of C4-substituted PYBOX ligand was designed and synthesized, and the optimal ligand was identified through systematic screening. Using CuCl2·2H2O/C4-substituted PyBox for the ACPA step, followed by AgOBz-catalyzed carboxylative cyclization under 2.0 MPa CO2 at 25 °C, a range of chiral 5-methylene-2-oxazolidinones were obtained in good yields with enantioselectivities up to 99% ee. The PYBOX ligand was found to be critical not only for the highly enantioselective propargylic amination step but also for facilitating the downstream AgOBz-catalyzed carboxylative cyclization process. The key to this one-pot tandem process was the use of triethylenetetramine as a chelating additive: it masks the upstream copper catalyst to prevent interference with the downstream cyclization step, while releasing the chiral ligand to activate the silver catalyst. Mechanistically, the copper-catalyzed propargylic amination generates a chiral propargylic amine, which subsequently undergoes silver-catalyzed carboxylative cyclization with CO2 to afford the target product. This work provided an efficient synthetic route to structurally diverse chiral 5-methylene-2-oxazolidinones and achieved, for the first time, the construction of such compounds bearing a tetrasubstituted stereocenter with high enantioselectivity. The synthetic utility was demonstrated through the total synthesis of (-)-cytoxazone and the formal synthesis of an ezetimibe analogue.
The hydrazine functional group, as an important fragment in drug molecules, exhibits biological activities distinct from those of amino groups. However, reports on the asymmetric construction of chiral oxazolidinones bearing a hydrazine moiety remain scarce. In 2025, building on their previous work, Zhou and co-workers further leveraged the newly developed chiral PYBOX ligands in combination with modifications to the C4 substituent of the pyridine ring to develop the first asymmetric propargylic hydrazination of ketone-derived propargylic esters (Scheme 24)[45]. On this basis, a CO2-involved asymmetric propargylic hydrazination-carboxylative cyclization tandem reaction was realized, enabling the highly enantioselective synthesis of chiral oxazolidinones bearing a hydrazine moiety and an α-tetrasubstituted carbon stereocenter. Single-crystal structural analysis revealed that the substituent at the C4 position of the pyridine ring influences the coordination ability of the PyBox ligand with copper salts. Furthermore, the chiral PyBox ligand was found to coordinate not only with copper salts but also with silver salts to form the corresponding complexes.
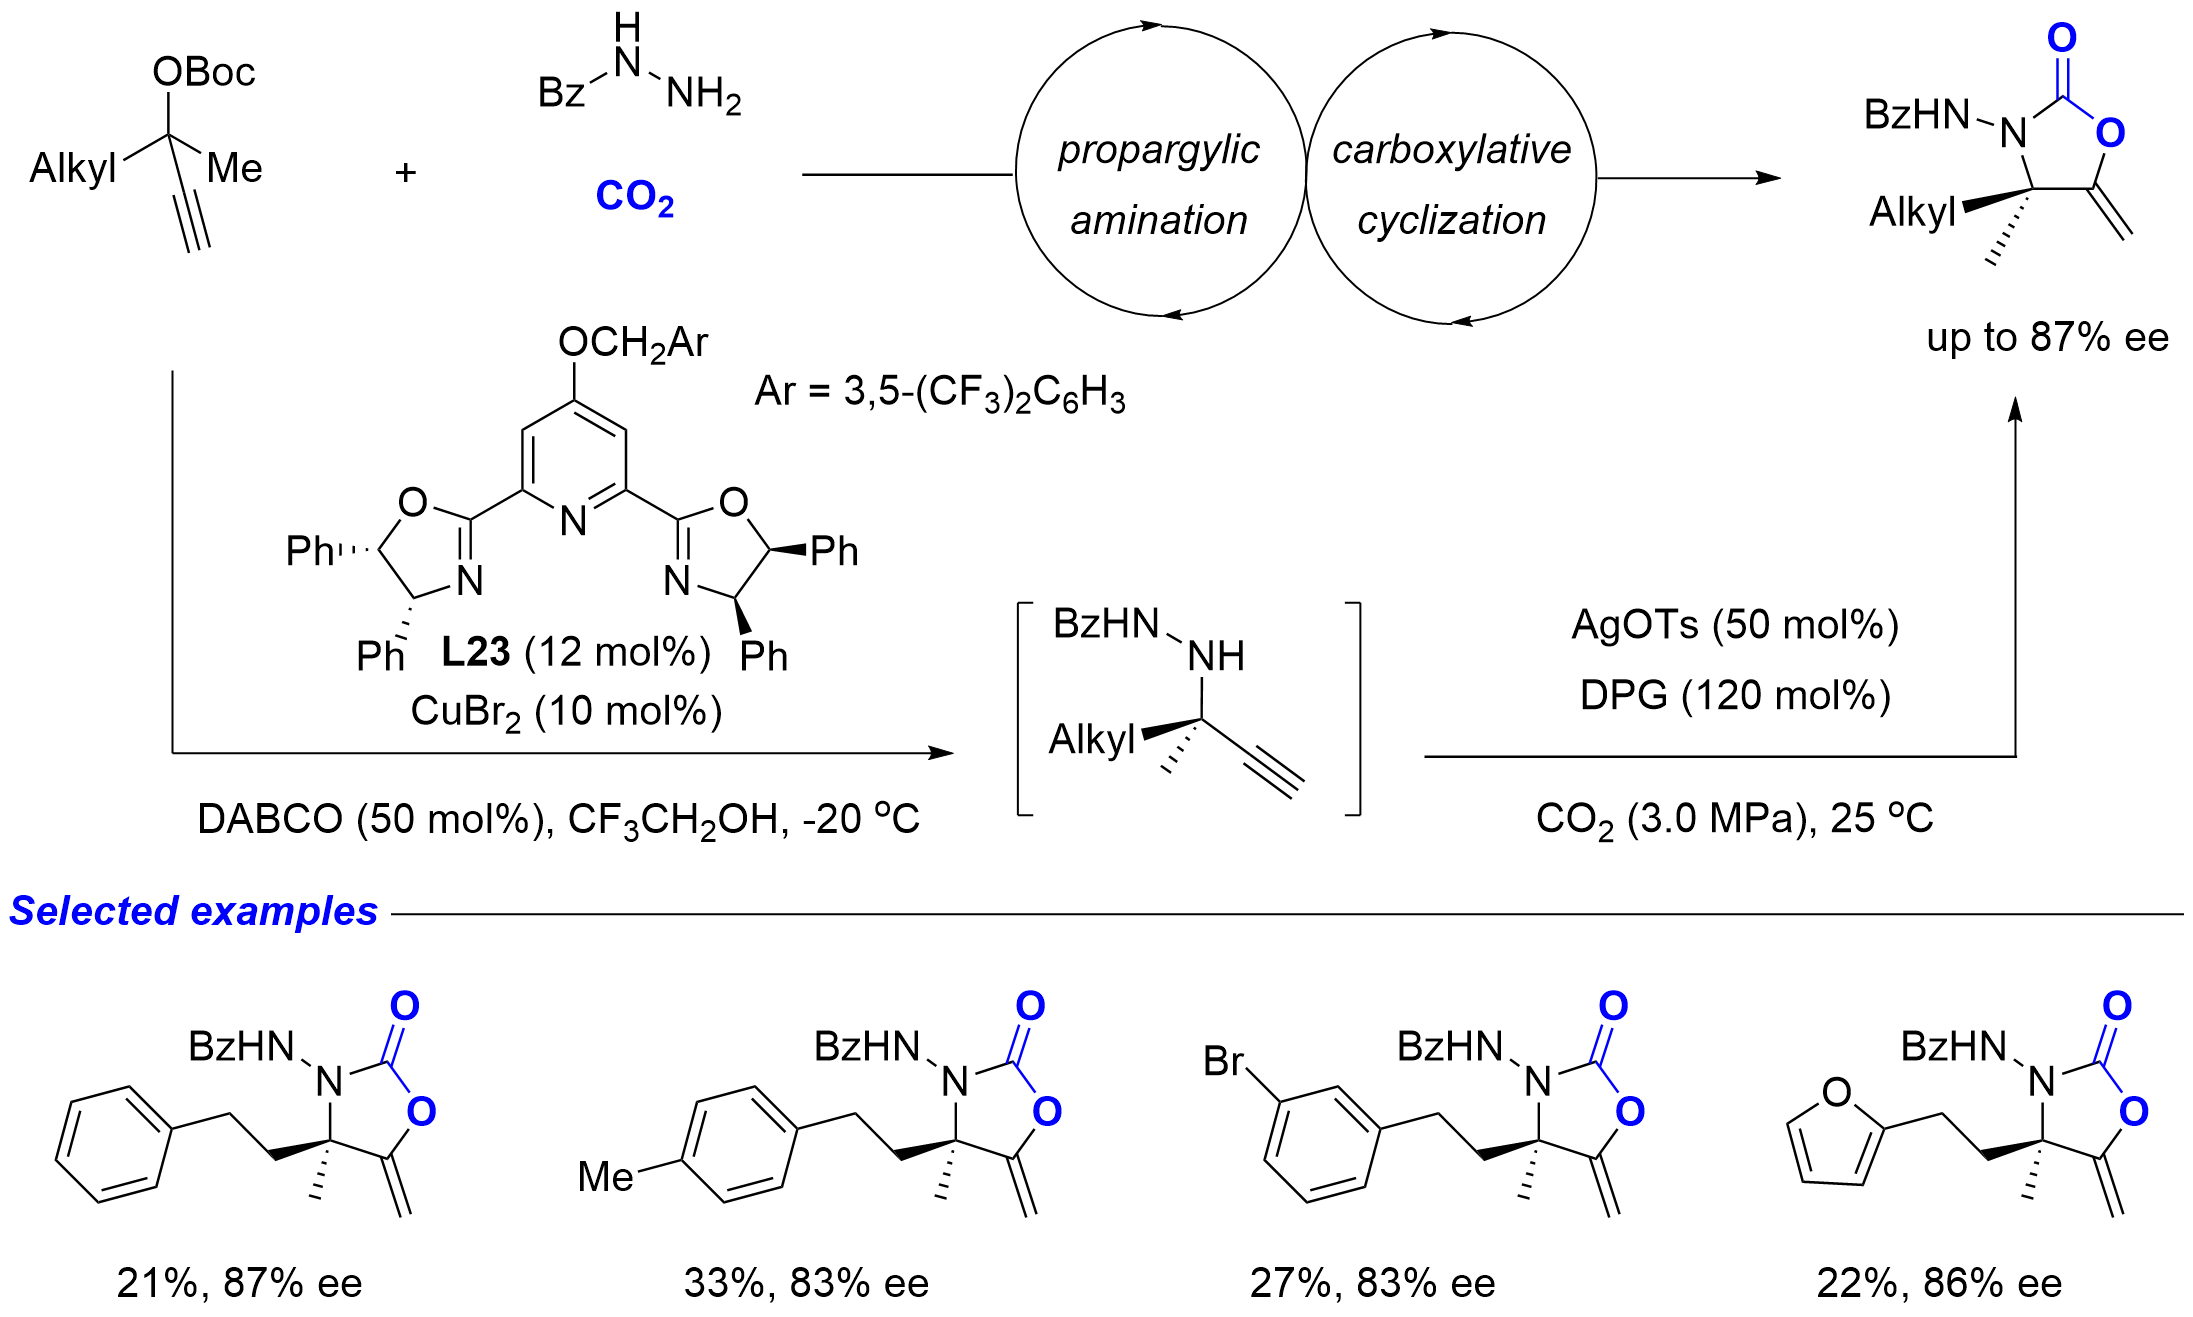
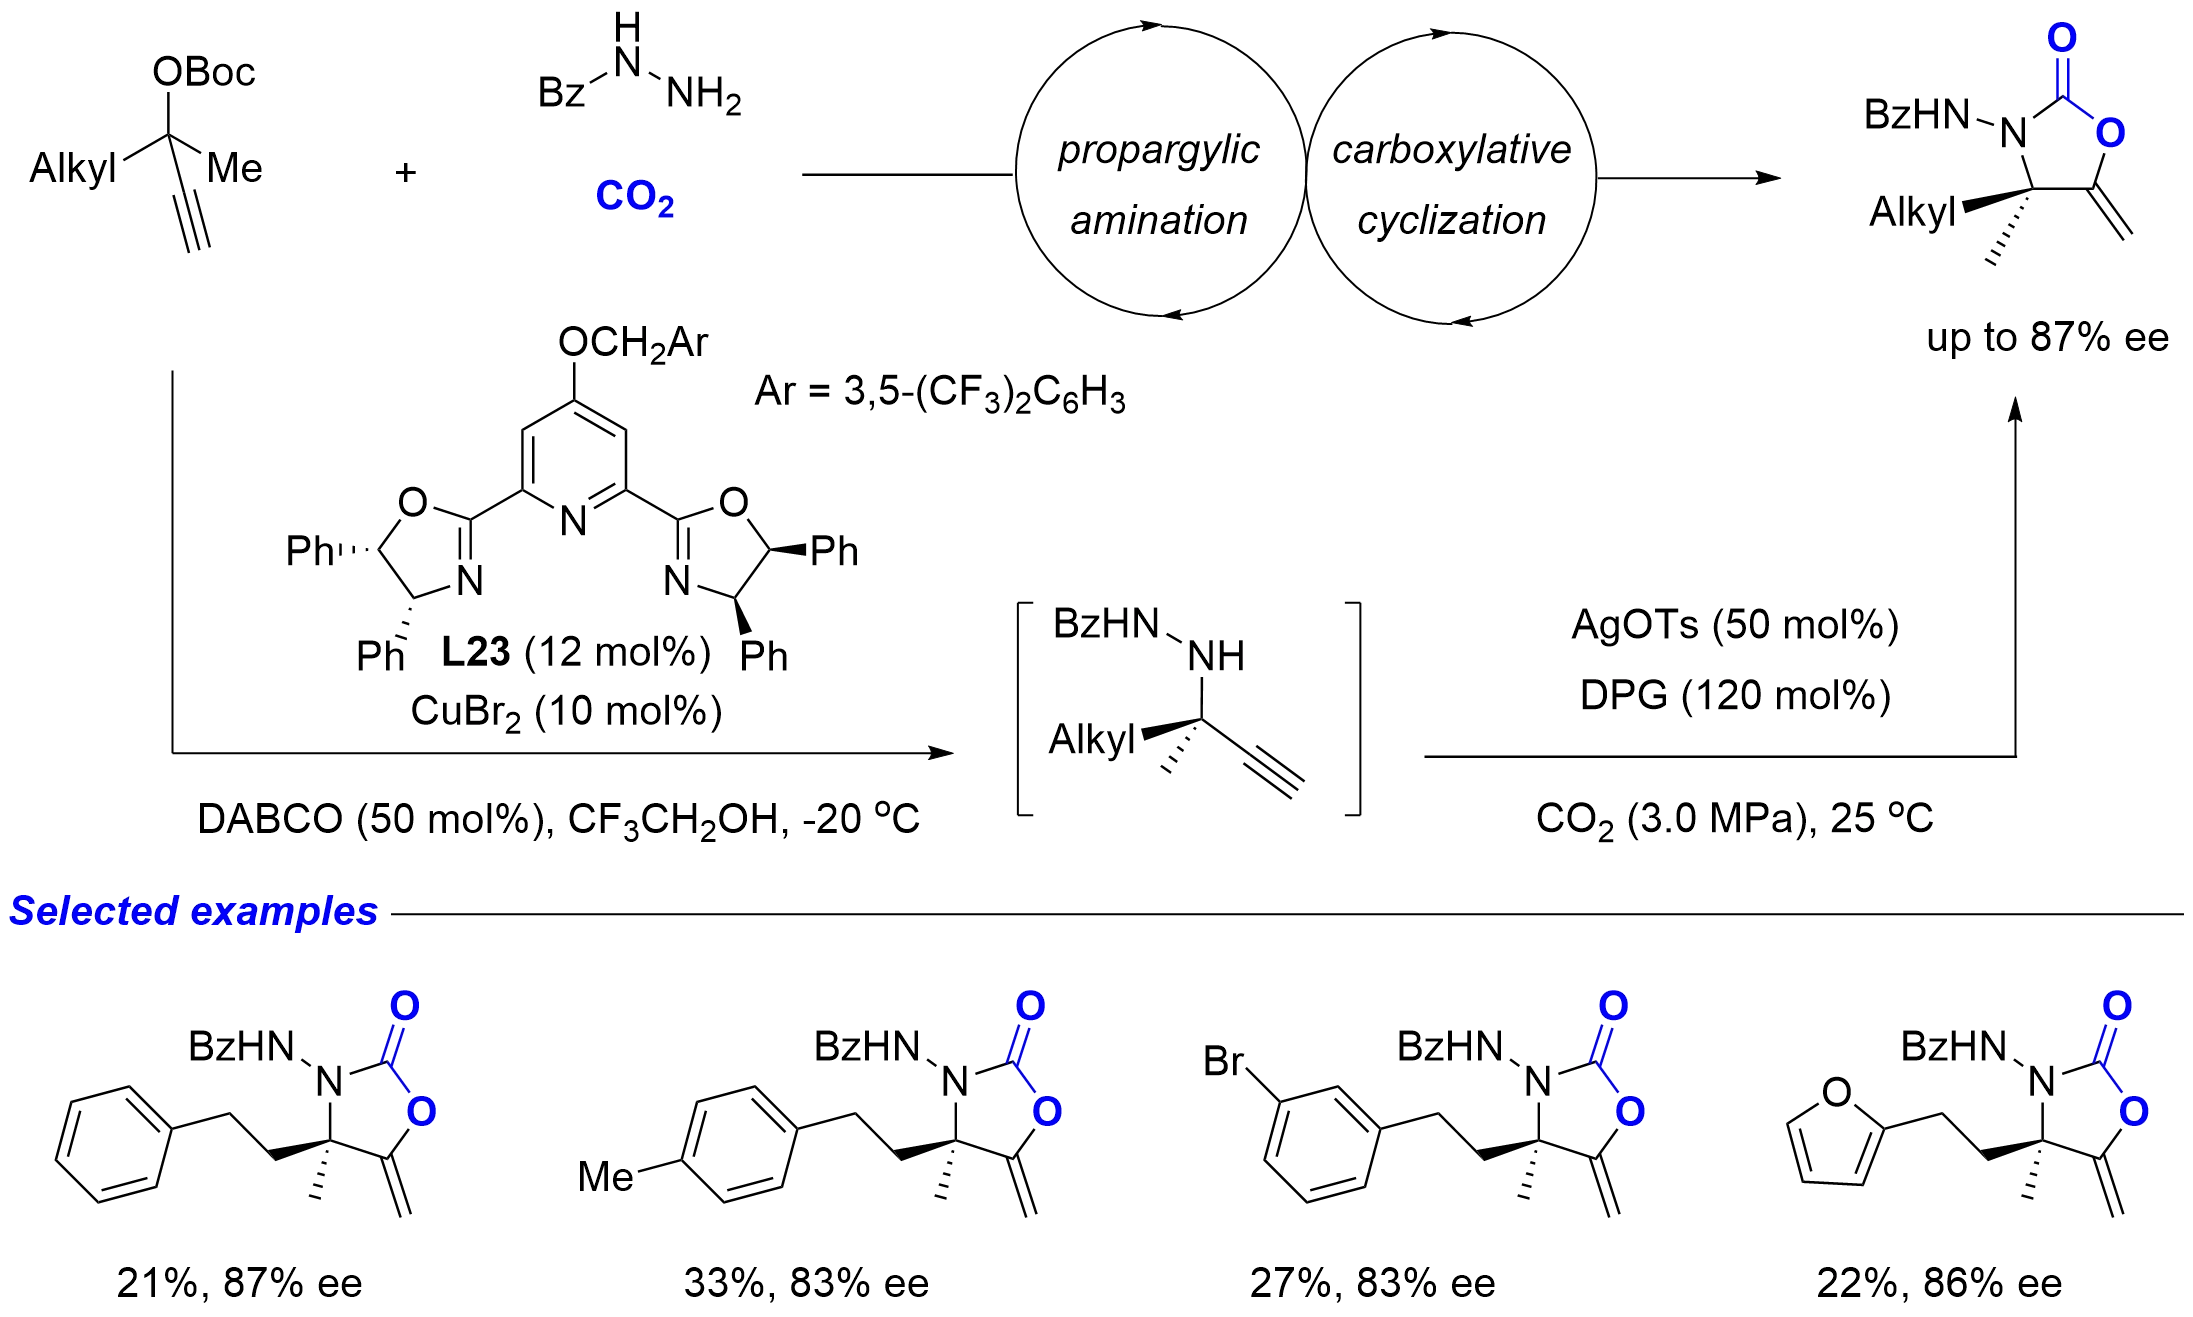
Scheme 24. Cu-catalyzed asymmetric propargylic hydrazination/carboxylative cyclization.
In addition to the above tandem catalytic strategies, asymmetric halocyclization has also provided a new approach for the construction of chiral 2-oxazolidinones. In 2023, Shirakawa and coworkers reported an asymmetric halocyclization reaction using chiral bifunctional chalcogenide catalysts (Scheme 25)[46]. Inspired by their previous work on catalytic asymmetric halolactonization, this strategy employs carbamic acids generated in situ from CO2 and allylamines as substrates, using a BINOL-derived chiral bifunctional selenide catalyst to achieve asymmetric halocyclization through an ordered transition state. The catalytic system exhibited good substrate generality for aryl-substituted allylamines, affording moderate to high enantioselectivities of up to 92% ee, although alkyl-substituted substrates gave lower selectivity. Mechanistic studies suggested that the catalyst first reacts with NBS to form a bromoselenonium intermediate. The hydroxy group of the catalyst then fixes the conformation of the carbamic acid substrate through hydrogen bonding with its carbonyl oxygen. Subsequent interaction of the alkene moiety with the bromoselenonium intermediate generates an ordered transition state, enabling stereoselective cyclization and catalyst regeneration.
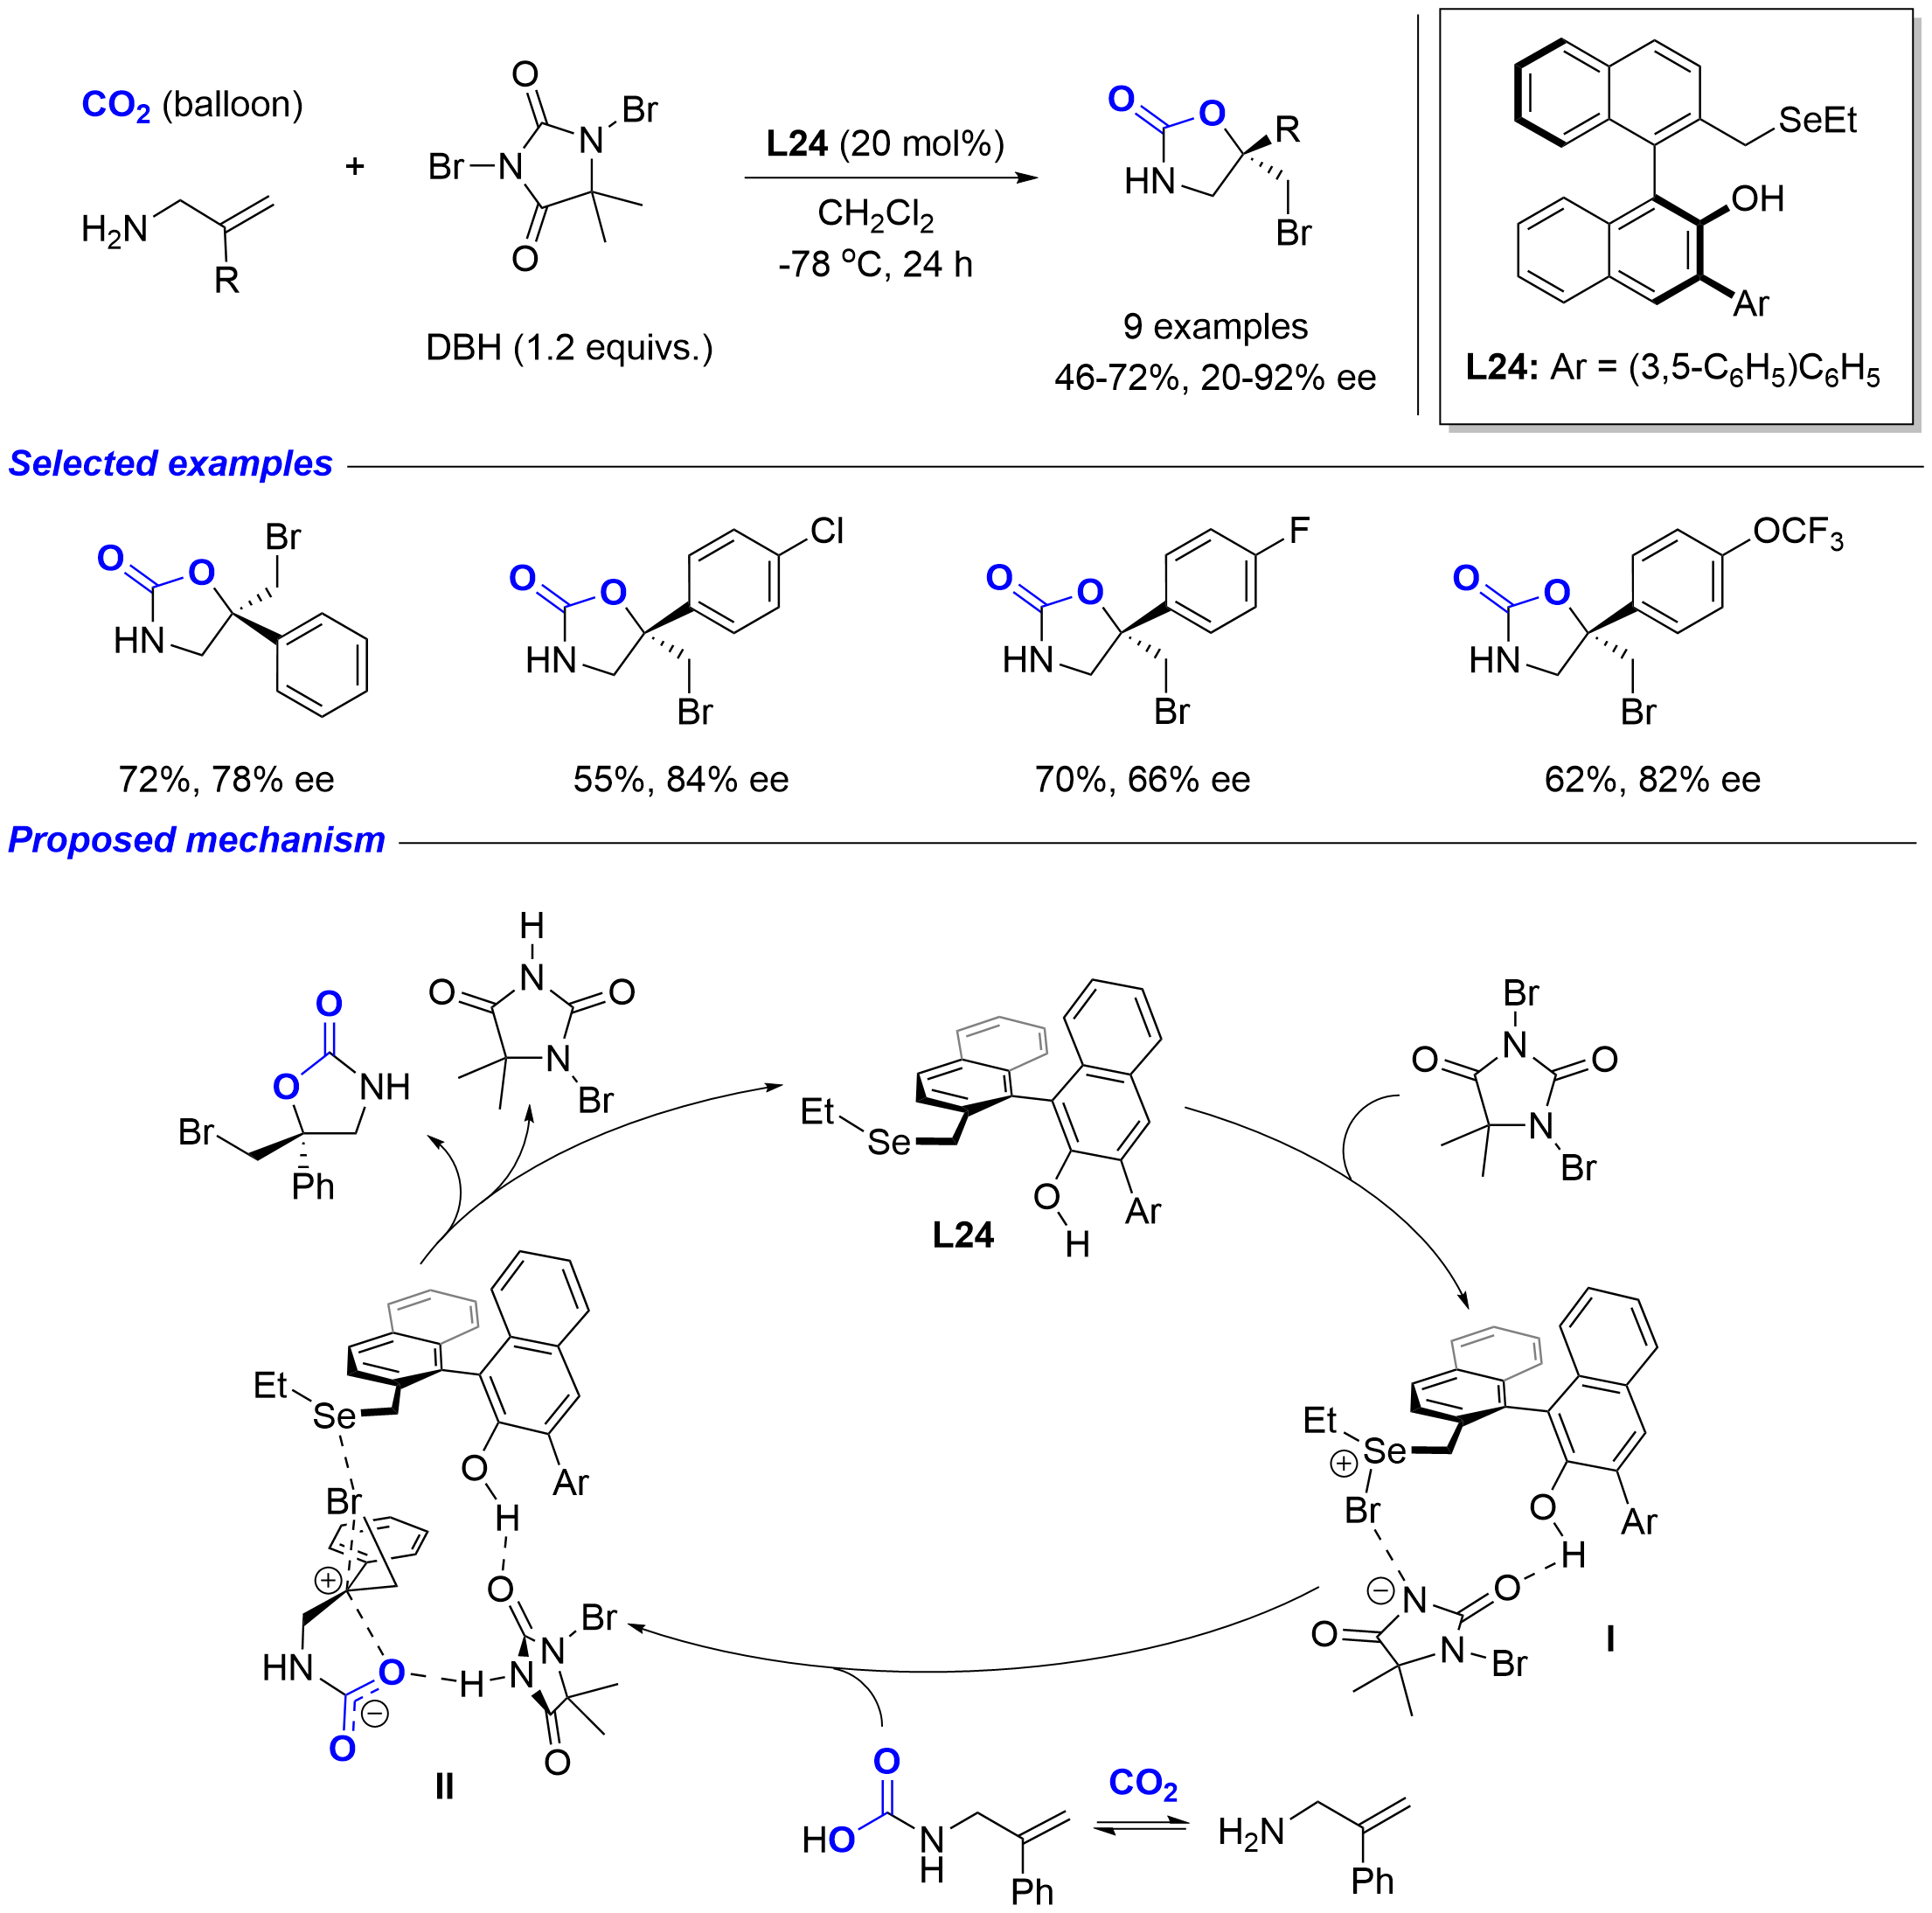
Scheme 25. Asymmetric bromocyclization of allylamines with CO2 to chiral 2-oxazolidinones.
Radical-mediated enantioselective C-O bond formation using CO2 as an oxygen source has also been explored. In 2025, Bao, Li, and co-workers reported an elegant copper-catalyzed radical enantioconvergent C-O bond-forming reaction, employing CO2 as the oxygen source to achieve asymmetric cross-coupling of allylamines with Togni reagent, thereby constructing chiral fluoroalkylated 2-oxazolidinone scaffolds (Scheme 26)[47]. The key to its success lies in the rational and strategic conversion of C2-symmetric bis(oxazoline) ligands to their C1-symmetric counterparts. Upon coordination with copper, these ligands induce additional C-N axial chirality, providing a well-defined chiral environment during the asymmetric C-O bond formation step, where the interaction between the radical intermediate and the metal center is relatively weak. Under optimized conditions, aryl-substituted allylamines afforded the corresponding fluoroalkylated 2-oxazolidinones in moderate to good yields with up to 92% ee.
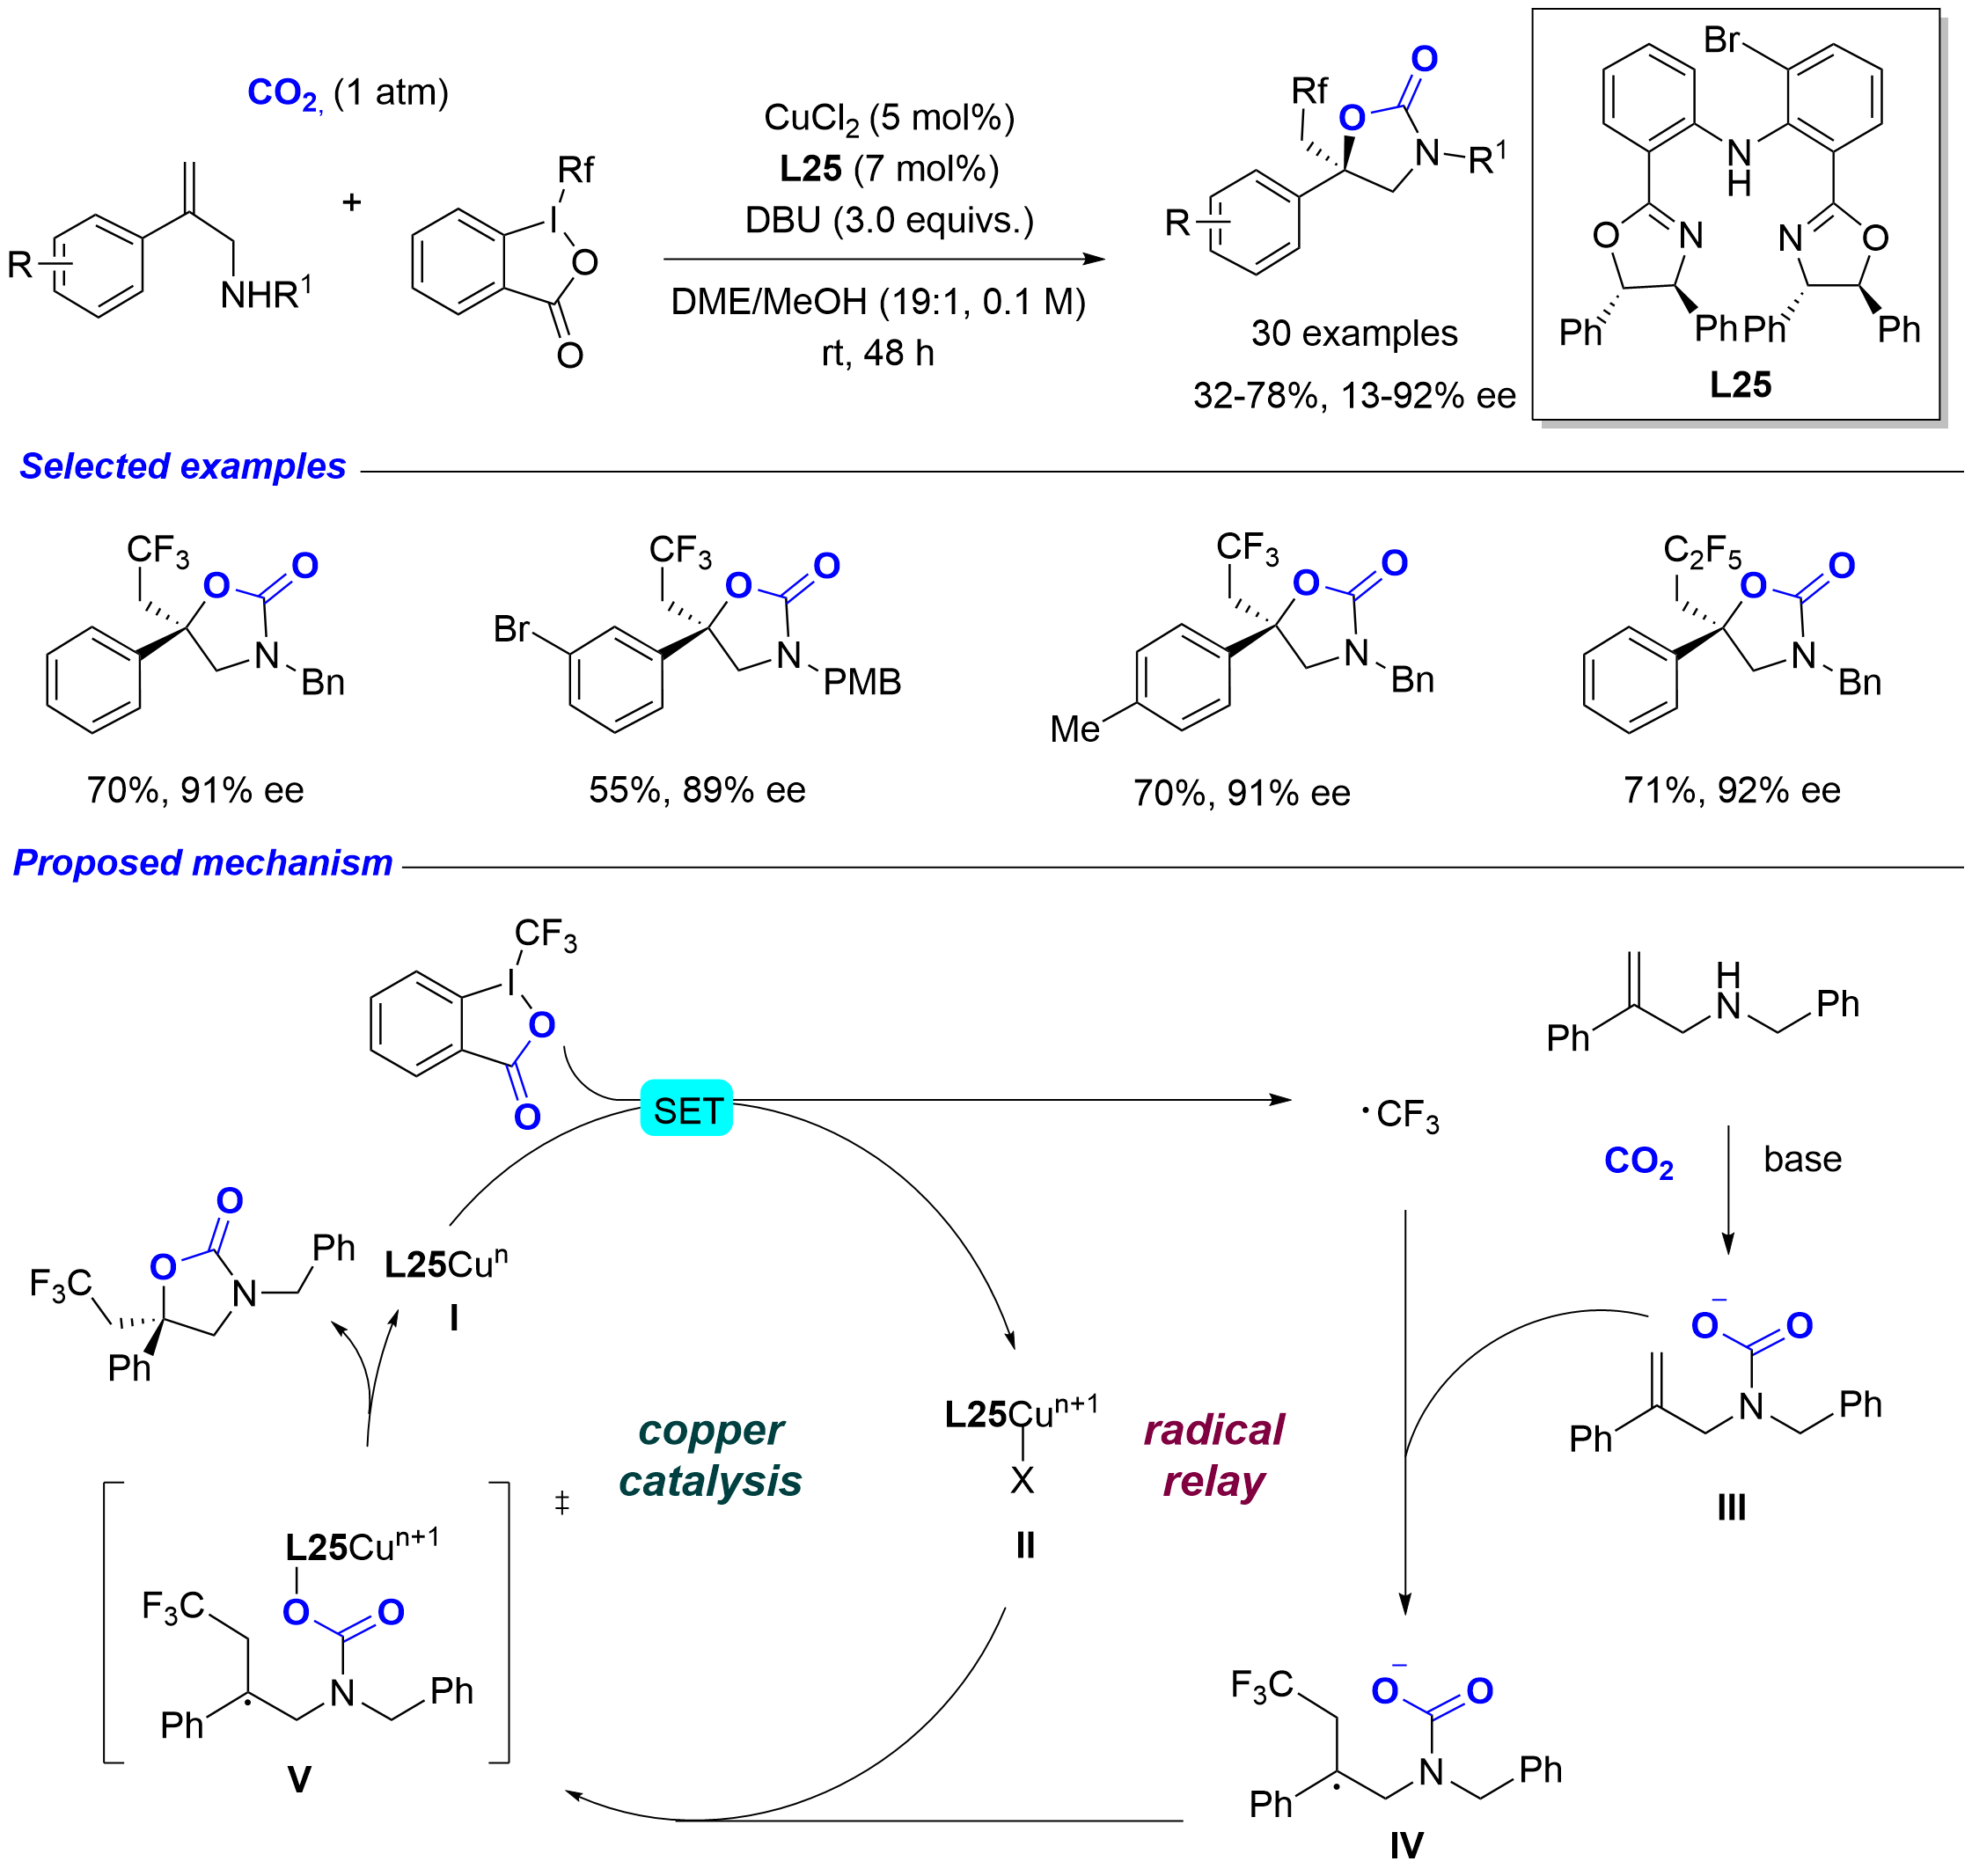
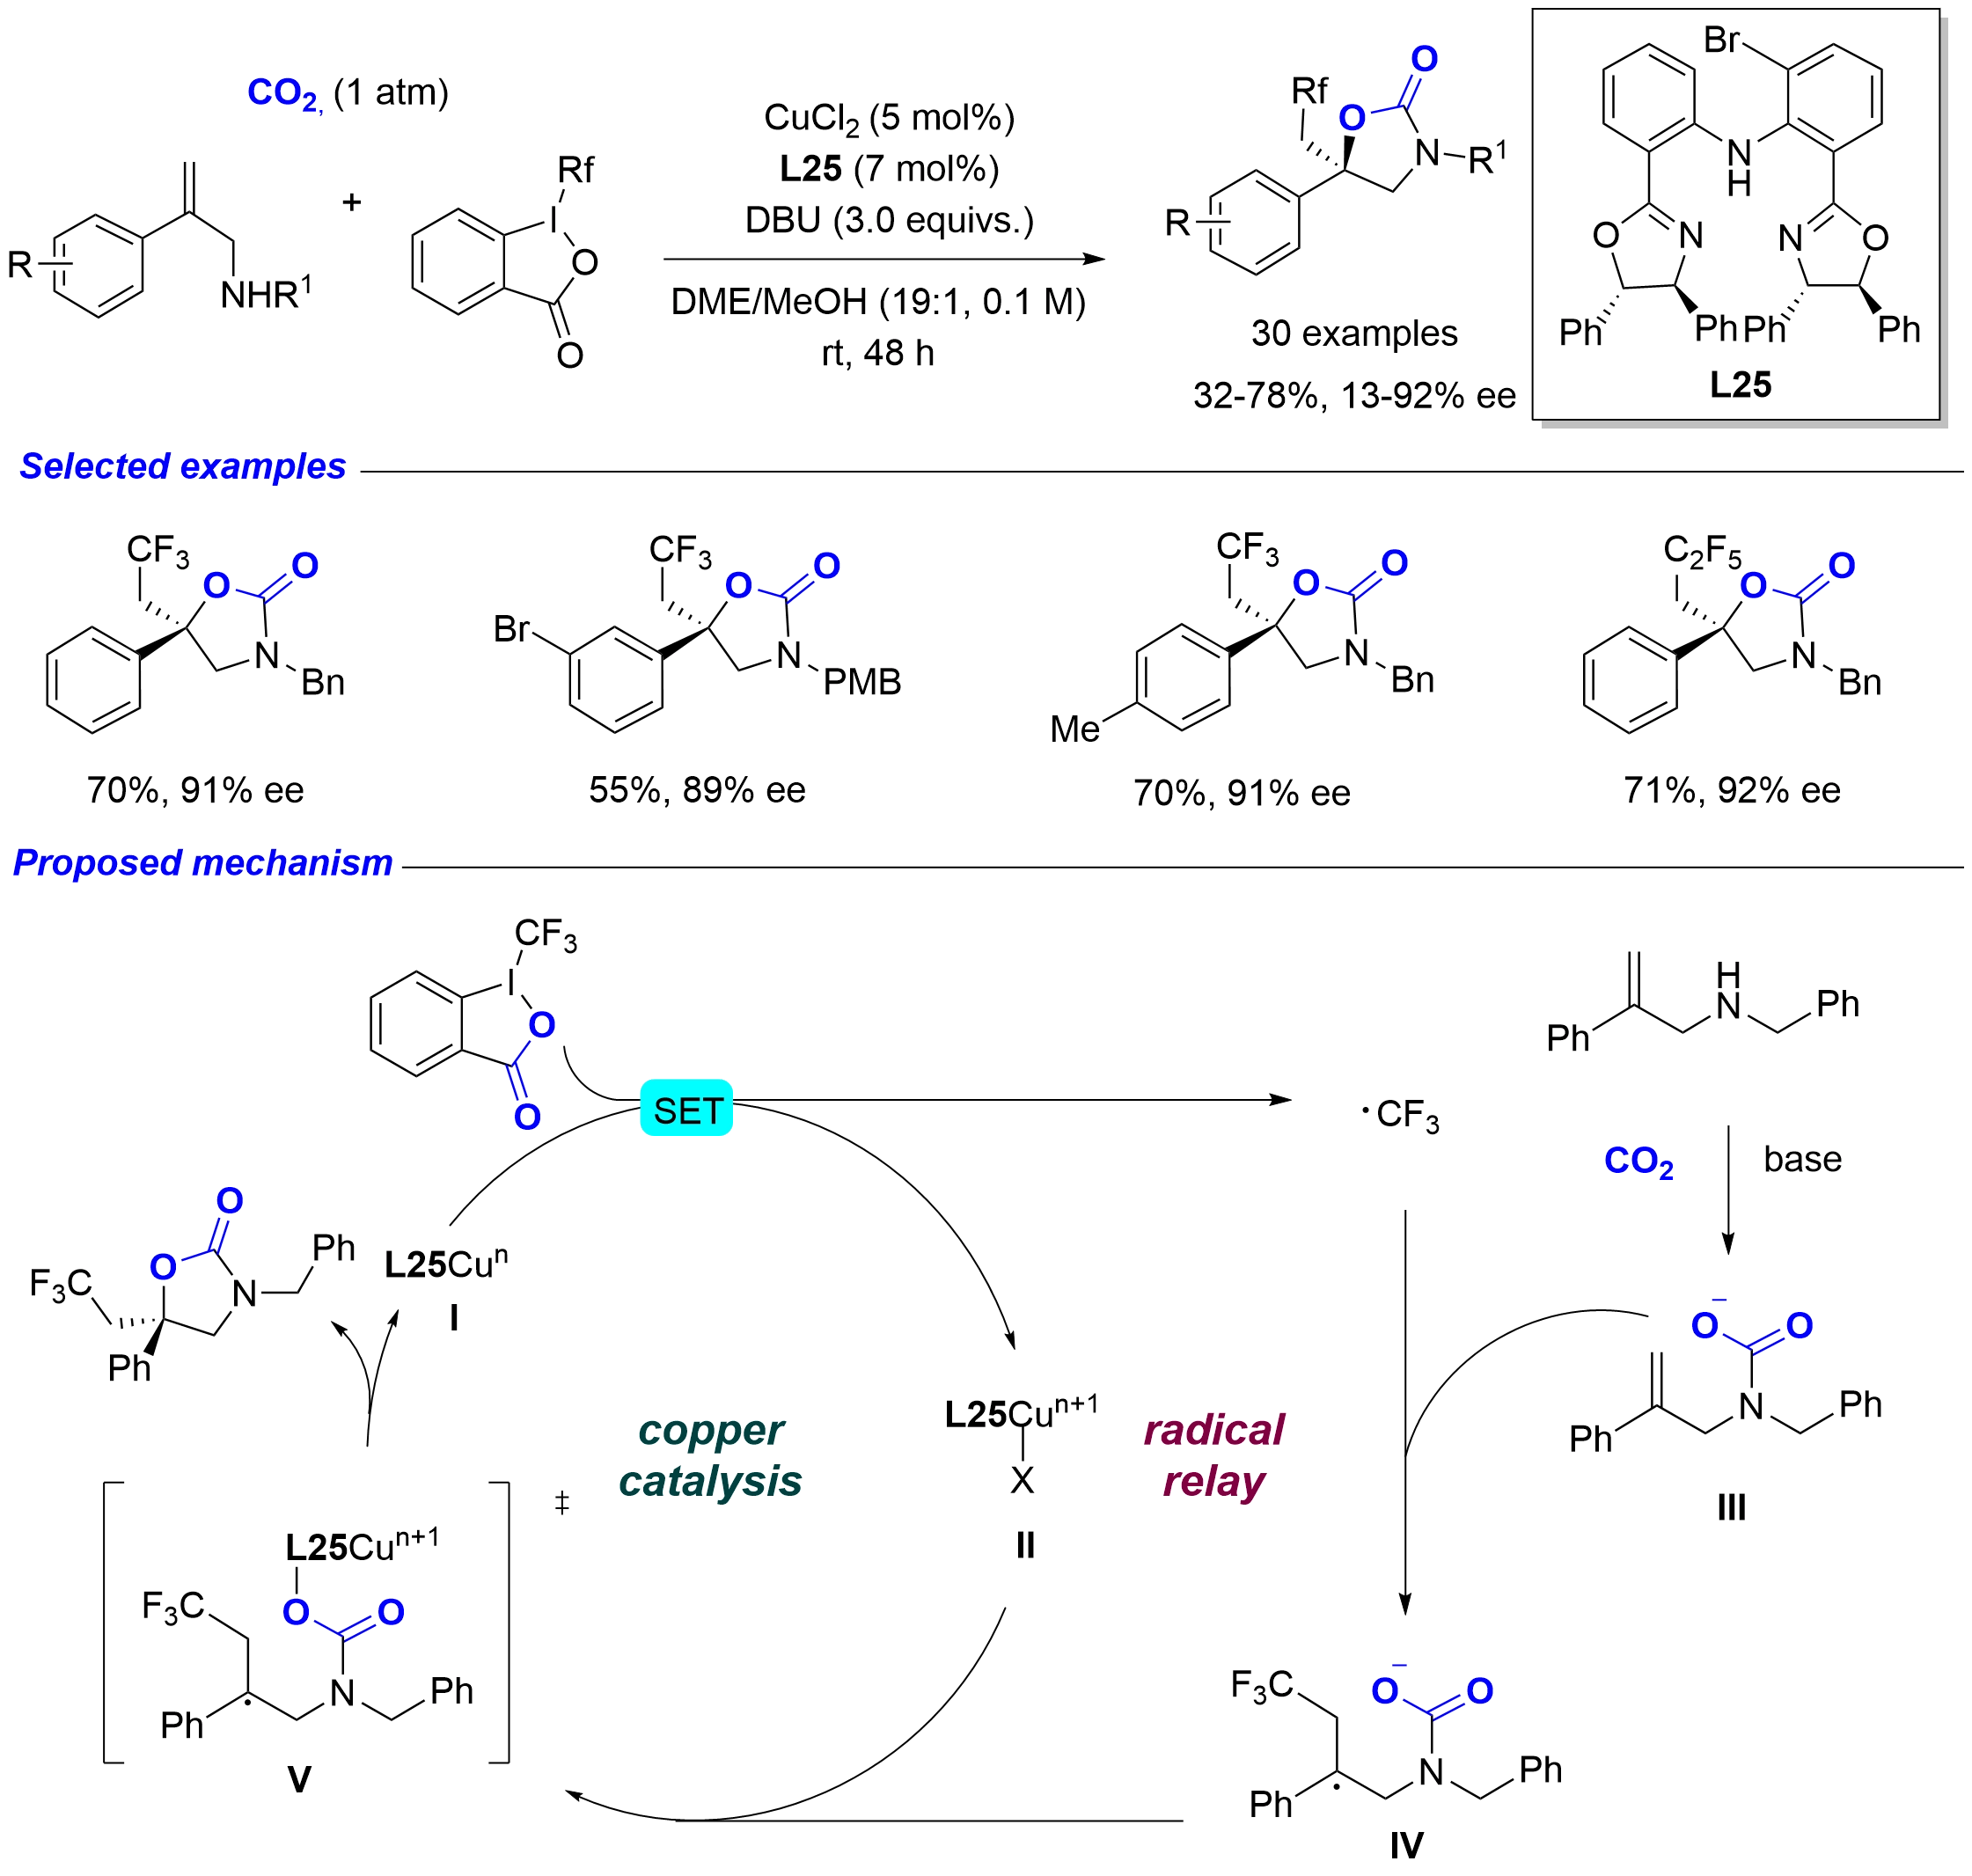
Scheme 26. Cu-catalyzed radical enantioconvergent C-O bond formation using CO2 as an oxygen source.
Mechanistic studies revealed that the C1-symmetric bisoxazoline ligand L25 derived chiral Cu complex undergoes in situ reduction to generate the active Cu species I. This species undergoes SET with Togni reagent to generate trifluoromethyl radical F3C• and a high-valent Cu intermediate II. Concurrently, under the action of the organic base DBU, allylamine condenses with CO2 to form an allylcarbamate anion intermediate III. Subsequently, F3C• undergoes radical addition to the alkene moiety of intermediate III, generating a carbon-centered radical intermediate IV, which coordinates with the Cu intermediate II to form a metal-radical complex V. Within the unique chiral cavity created by the C1-symmetric ligand, complex V undergoes either a group transfer or reductive elimination to achieve stereoselective C-O bond formation, furnishing the chiral fluoroalkylated 2-oxazolidinone product and regenerating the Cu species I to complete the catalytic cycle.
3.2 Asymmetric synthesis of chiral cyclic carbonates with CO2
Chiral cyclic carbonates constitute an important class of chiral heterocyclic compounds. The most atom-economical route to their synthesis is the direct asymmetric cycloaddition of epoxides with CO2[16,17]. The success of this transformation hinges on the design of chiral catalysts that can simultaneously activate the epoxide, control the stereoselectivity of nucleophilic ring-opening, and efficiently capture CO2.
In 2021, Larionov and coworkers reported the enantioselective kinetic resolution of epoxides coupled with CO2 cycloaddition, catalyzed by octahedral Co(III) complexes bearing metal-centered chirality derived from (R, R)-1,2-diphenylethylenediamine, achieving a selectivity factor of up to 2.8 (Scheme 27)[48]. In 2022, the same group extended this strategy using similar catalytic systems and reported s values of up to 1.6[49]. Kinetic experiments and DFT calculations revealed a first-order dependence on catalyst concentration, identified epoxide ring-opening as the rate-determining step, and elucidated a catalytic cycle comprising hydrogen-bond donor activation, nucleophilic ring-opening by iodide, CO2 insertion, and cyclization. Later in 2024, they further reported a chiral Ni(II)-catalyzed kinetic resolution of epoxides with CO2, achieving a selectivity factor of up to 11.5 in the presence of a co-ligand[50]. Mechanistic studies revealed that the Ni(II) species (S)-Ni and the co-ligand (S)-L26 assemble into an octahedral structure, activating the epoxide through dual hydrogen bonding, with nucleophilic ring-opening by iodide serving as the stereodetermining step.
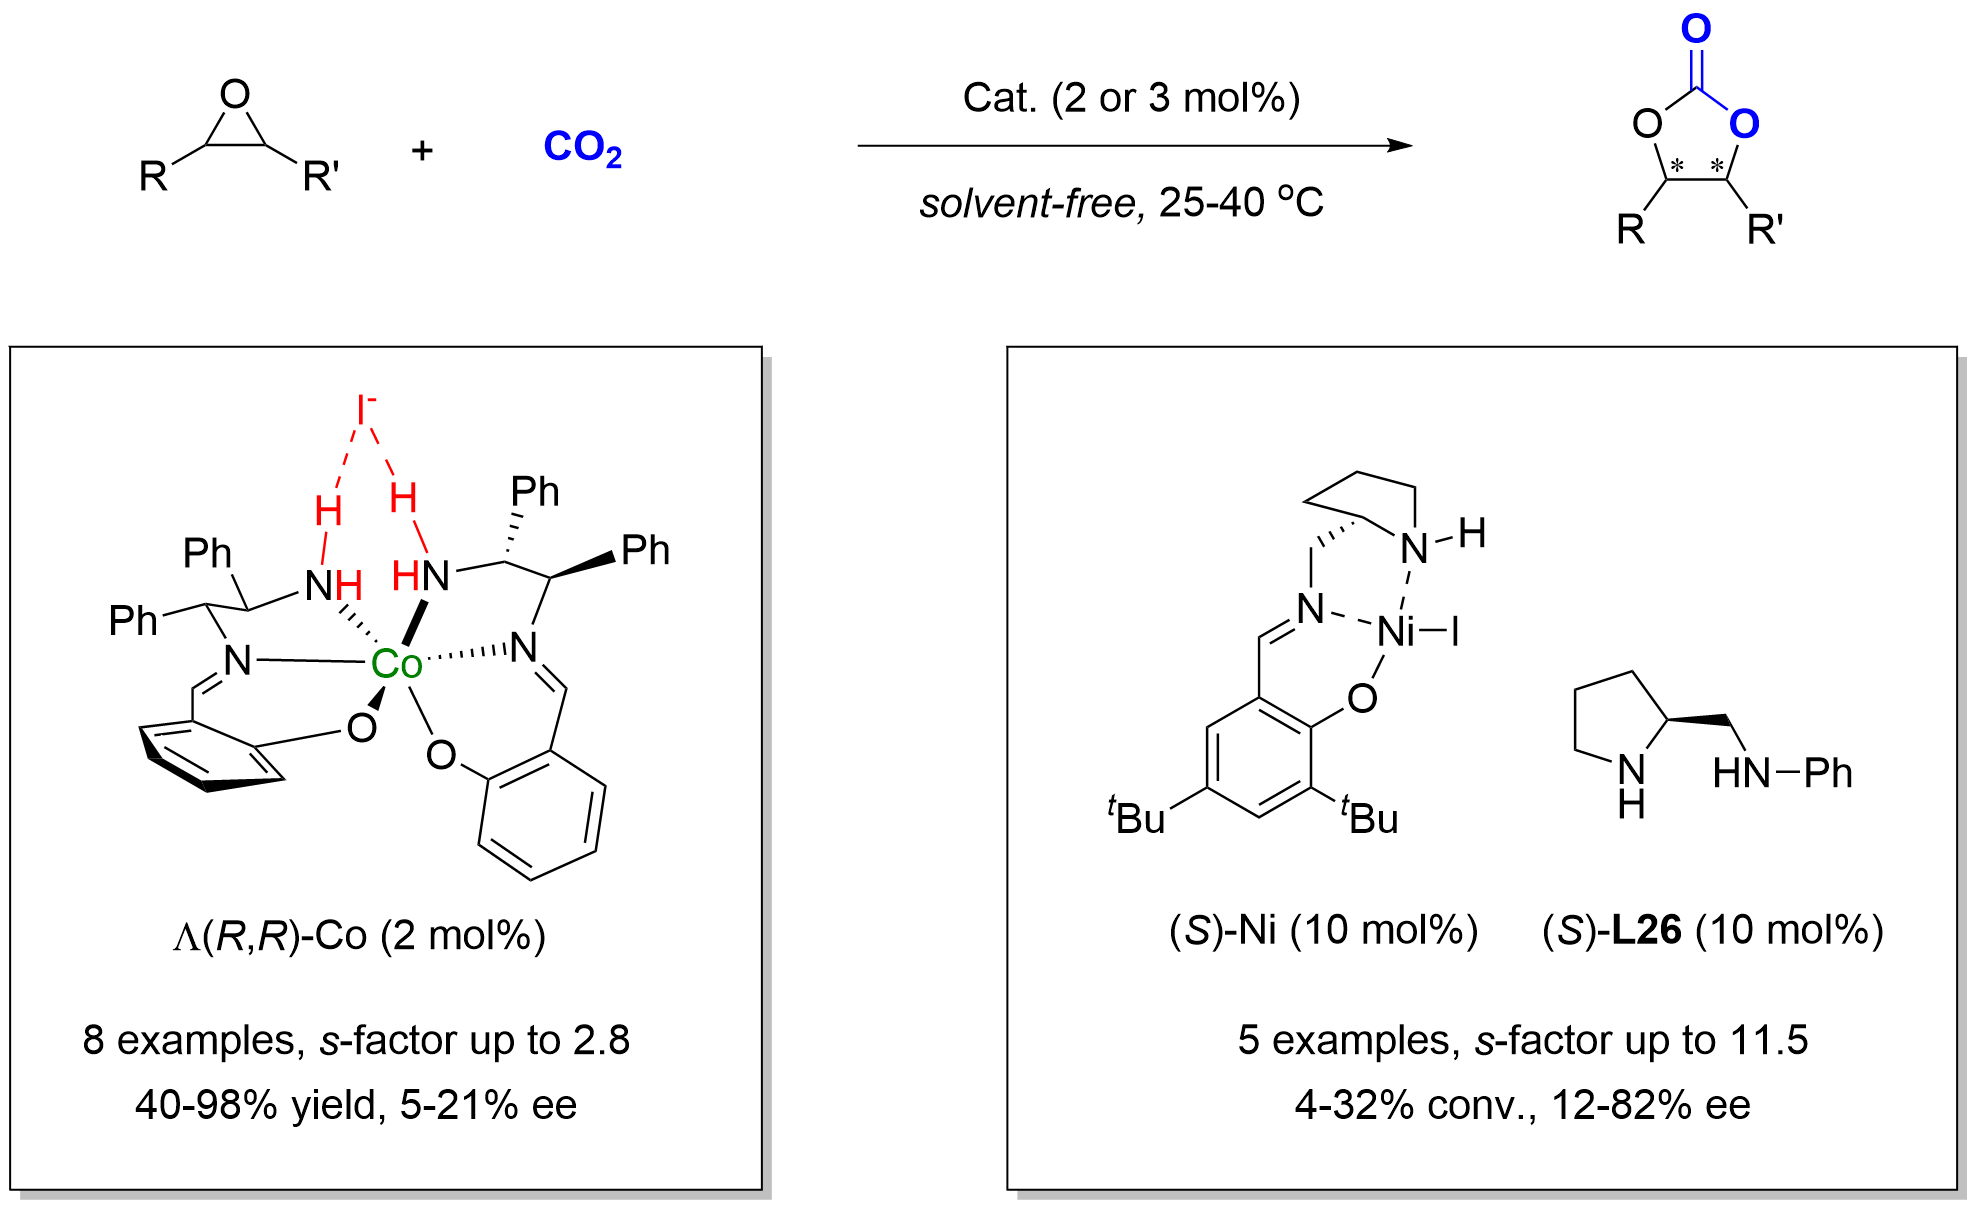
Scheme 27. Chiral metal-catalyzed kinetic resolution and cycloaddition of epoxides with CO2.
In addition to the aforementioned catalytic systems, Esteve and García-Verdugo, inspired by the synergistic supramolecular interactions in enzymatic catalysis, designed a class of dinuclear Zn(II) complexes based on pseudopeptidic macrocyclic ligands for the cycloaddition of CO2 with epoxides, enabling the synthesis of chiral cyclic carbonates (Scheme 28)[51]. These catalysts feature a preorganized cavity, in which the Zn(II) centers and iodide ions cooperatively activate the substrates without the need for an external cocatalyst. Under 1 bar CO2 at 80 °C, styrene oxide was converted quantitatively with a TOF of 198 h-1. Through structural modulation at the chiral amino acid side chains, this system enabled the kinetic resolution of racemic epoxides, with the phenylalanine-derived catalyst exhibiting a selectivity factor of 9.5 for styrene oxide. DFT calculations revealed that non-covalent interactions, including hydrogen bonding and π-π stacking, participate in substrate recognition and stereoselectivity control.
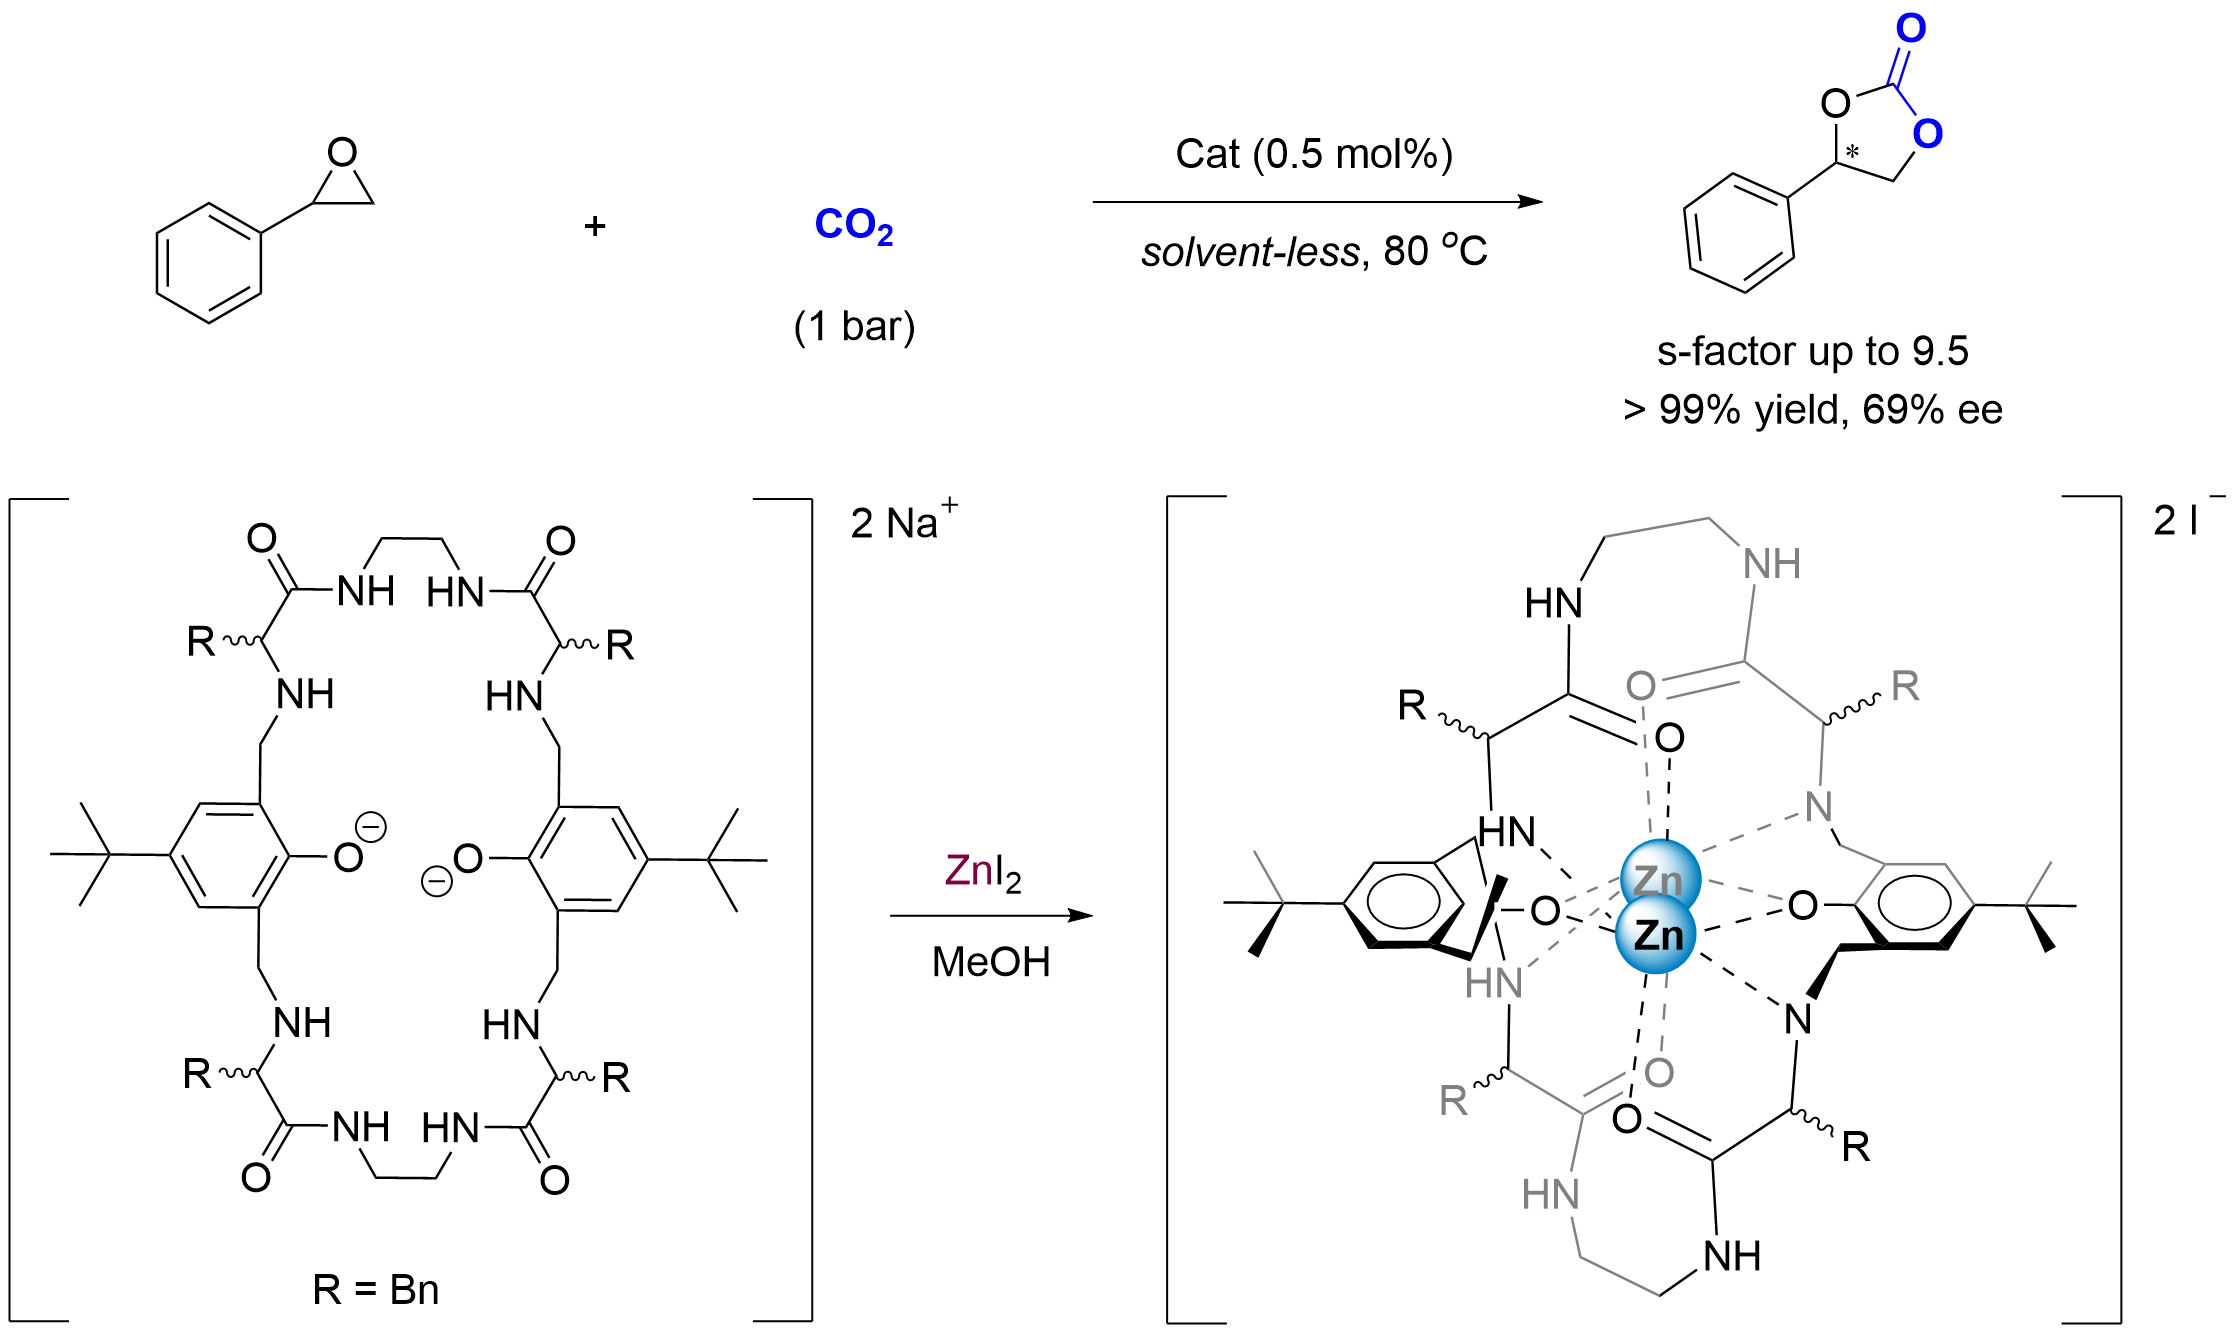
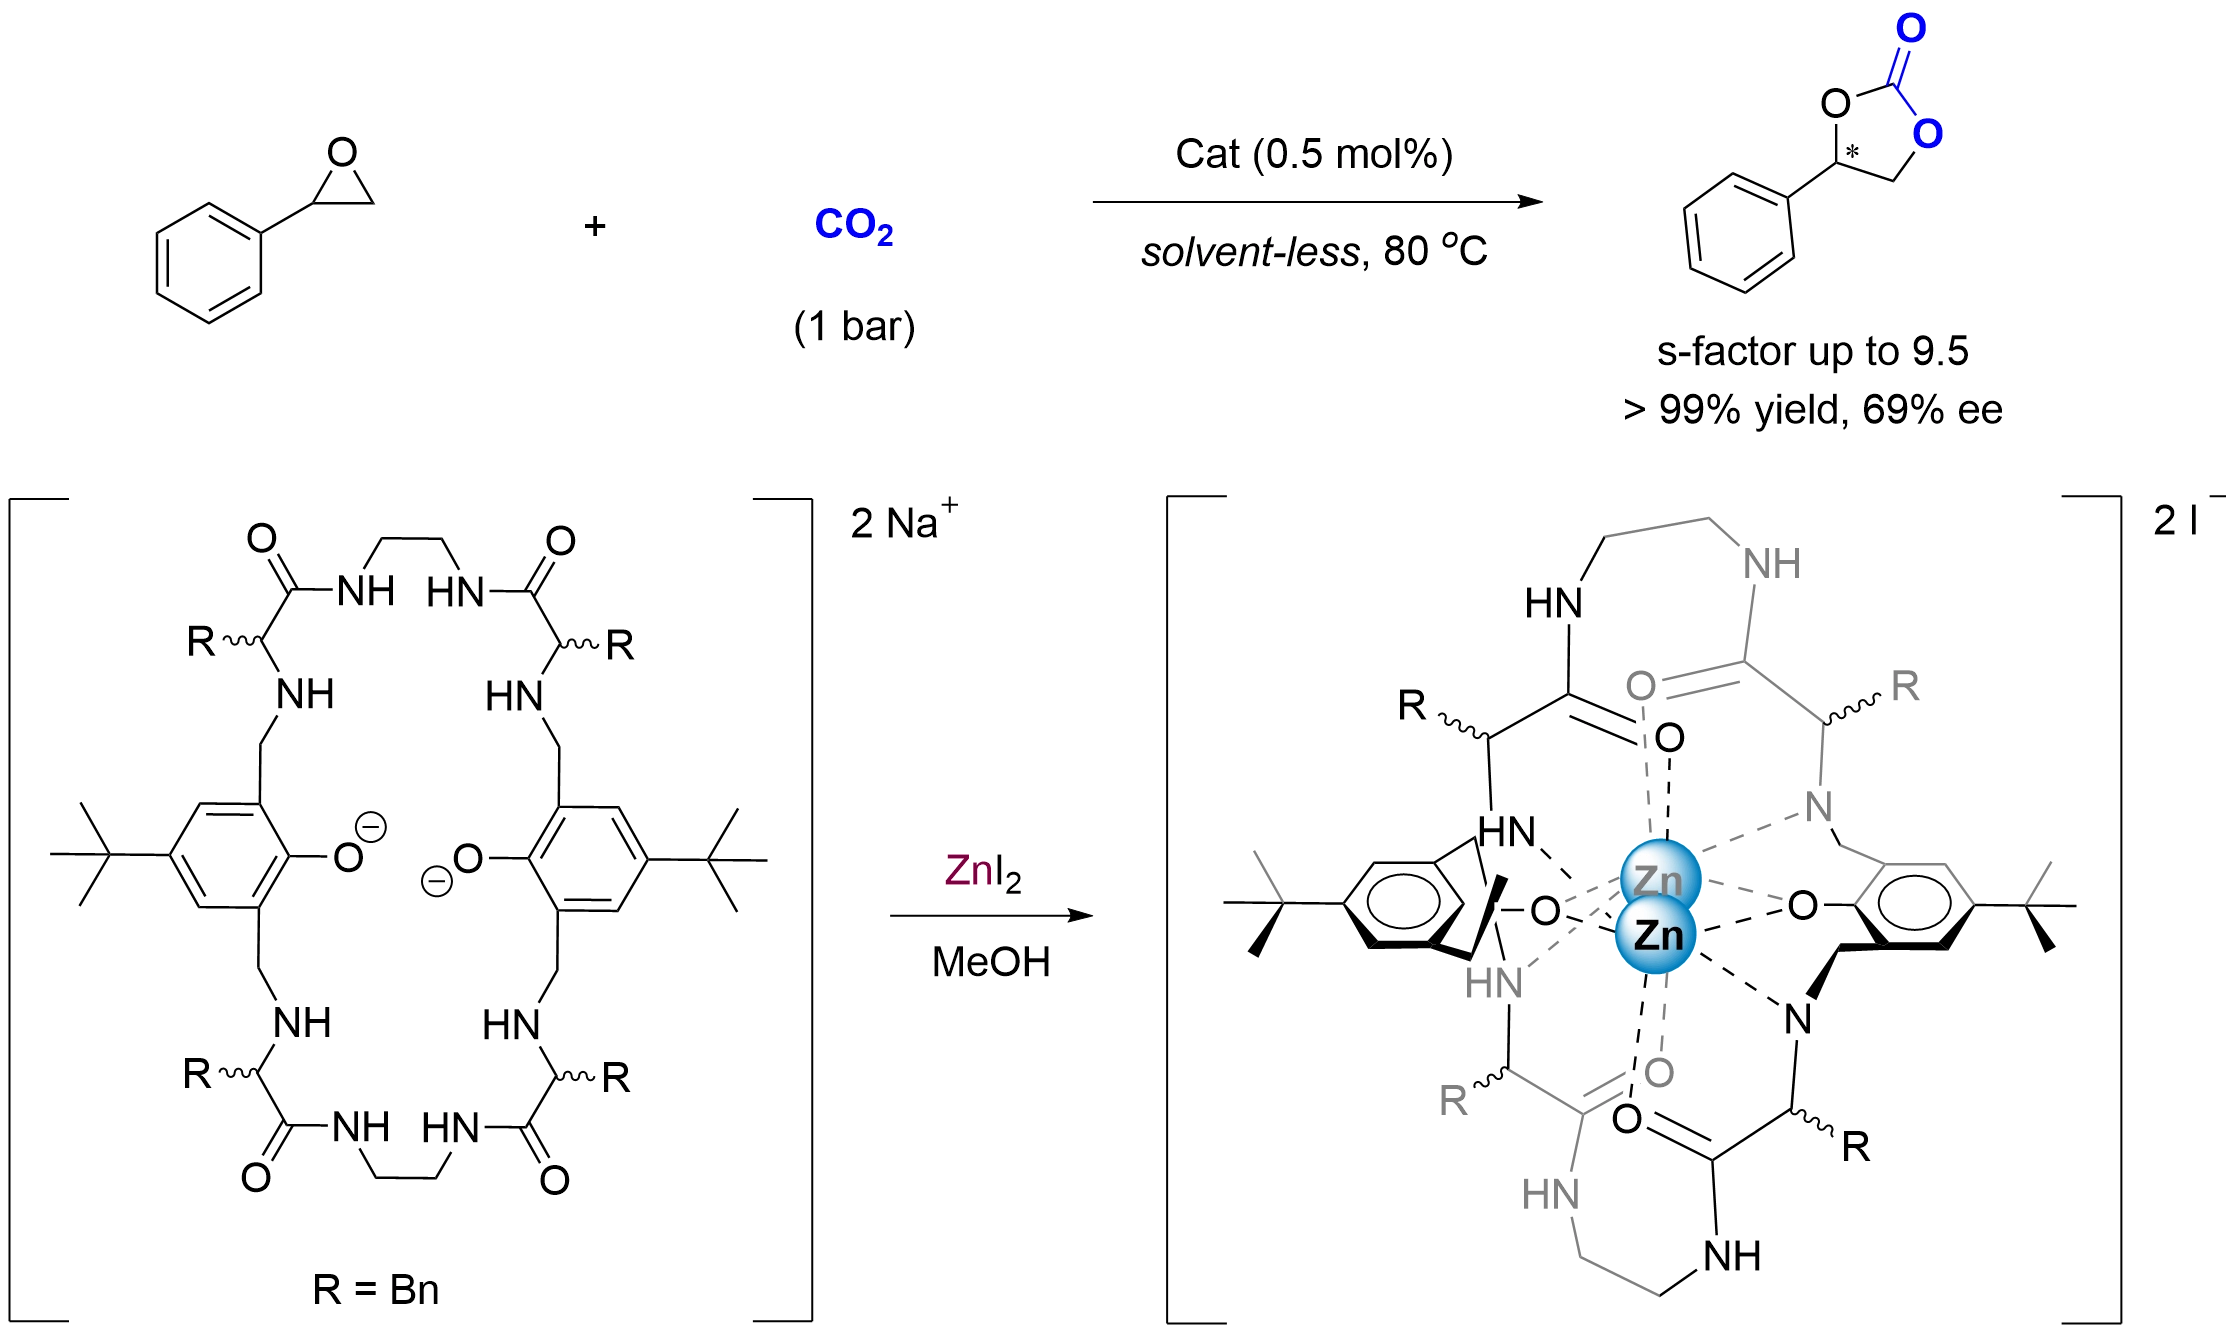
Scheme 28. Pseudopeptidic dinuclear Zn(II)-catalyzed asymmetric cycloaddition of CO2 to epoxides with kinetic resolution.
4. Summary and Outlook
Significant progress has been made in the asymmetric synthesis of chiral carboxylic acids and chiral heterocycles using CO2 as a C1 synthon. For chiral carboxylic acids, strategies including carbo-carboxylation, boracarboxylation, hydrocarboxylation, dicarboxylation, and electroreductive carboxylation have been developed using carbon-carbon unsaturated bonds and organic electrophiles, affording products with central or axial chirality. Several methods have been applied to the total synthesis of natural products and pharmaceuticals. In chiral heterocycle synthesis, representative targets such as 2-oxazolidinones and cyclic carbonates have been constructed via tandem carboxylative cyclization, radical enantioconvergent C-O bond formation, and cycloaddition. These advances underscore the potential of CO2 as a green and sustainable C1 resource in chiral fine chemical synthesis.
Despite this progress, considerable challenges remain. (i) Current catalytic systems rely heavily on transition metals; research on metal-free organocatalysts remains limited. (ii) Most methods require strictly anhydrous and anaerobic conditions or relatively high CO2 pressures; developing efficient asymmetric transformations under ambient conditions or with low-concentration CO2 is a key step toward practical implementation. (iii) While excellent enantioselectivities (> 95% ee) have been achieved for activated substrates such as aryl alkenes and aryl halides, simultaneous control of regio-, enantio-, and diastereoselectivity for unactivated alkyl alkenes, alkyl-substituted alkynes, and complex scaffolds with multiple stereocenters remains challenging. (iv) Most existing asymmetric transformations are centered on highly reactive substrates with pre-existing functional groups or activated bonds. In contrast, asymmetric catalytic transformations involving CO2 with inert chemical bonds, such as unactivated C(sp3)-H bonds, C-C σ-bonds, and C-O σ-bonds, remain largely unexplored[52]. (v) Most studies remain at the methodological stage, with limited application in late-stage modification and scalable synthesis of complex drug molecules, natural products, and functional materials. Greater emphasis should be placed on integrating CO2-based asymmetric catalysis with total synthesis, chiral ligand design, and bulk chemical upgrading, as well as compatibility with continuous flow, scale-up, and biomass-derived substrates.
We believe that continued innovation in catalytic systems, reaction modes, and chiral control strategies, combined with integration of electrochemistry[53,54], photochemistry[55,56], flow chemistry[57], and AI-assisted design, will drive further progress in asymmetric catalysis involving CO2 and build a stronger bridge between green chemistry and chiral synthesis[58].
Authors contribution
Yang D, Zhou H, Wang L, Wang H, Jiang M, Zhang Y, Chen D, Pan X: Investigation, data curation, visualization, writing-original draft.
Zhou J: Conceptualization, methodology.
Zhou F: Conceptualization, supervision, project administration, writing-review & editing.
Conflicts of interest
The authors declare no competing interests.
Ethical approval
Not applicable.
Consent to participate
Not applicable.
Consent for publication
Not applicable.
Availability of data and materials
Not applicable.
Funding
The financial support from the National Natural Science Foundation of China (Grant Nos. 22571092 and 22171090), the National Key Research and Development Program of China (Grant No. 2020YFA0710200), and the Innovation Program of Shanghai Municipal Education Commission (2023ZKZD37) is highly appreciated.
Copyright
© The Author(s) 2026.
References
-
1. (a) Sakakura T, Choi JC, Yasuda H. Transformation of carbon dioxide. Chem Rev. 2007;107(6):2365-2387.[DOI](b) Tsuji Y, Fujihara T. Carbon dioxide as a carbon source in organic transformation: Carbon-carbon bond forming reactions by transition-metal catalysts. Chem Commun. 2012;48(80):9956-9964.[DOI](c) He M, Sun Y, Han B. Green carbon science: Scientific basis for integrating carbon resource processing, utilization, and recycling. Angew Chem Int Ed. 2013;52(37):9620-9633.[DOI](d) Aresta M, Dibenedetto A, Angelini A. Catalysis for the valorization of exhaust carbon: From CO2 to chemicals, materials, and fuels. Technological use of CO2. Chem Rev. 2014;114(3):1709-1742.[DOI](e) Fiorani G, Wusheng G, Kleij AW. Sustainable conversion of carbon dioxide: The advent of organocatalysis. Green Chem. 2015;17(3):1375-1389.[DOI](f) Sanz-Pérez ES, Murdock CR, Didas SA, Jones CW. Direct capture of CO2 from ambient air. Chem Rev. 2016;116(19):11840-11876.[DOI](g) Artz J, Müller TE, Thenert K, Kleinekorte J, Meys R, Sternberg A, et al. Sustainable conversion of carbon dioxide: An integrated review of catalysis and life cycle assessment. Chem Rev. 2018;118(2):434-504.[DOI](h) Burkart MD, Hazari N, Tway CL, Zeitler EL. Opportunities and challenges for catalysis in carbon dioxide utilization. ACS Catal. 2019;9(9):7937-7956.[DOI](i) Dibenedetto A, Nocito F. The future of carbon dioxide chemistry. ChemSusChem. 2020;13(23):6219-6228.[DOI]
-
2. (a) Aresta M. Carbon dioxide as chemical feedstock. Weinheim: Wiley-VCH Verlag GmbH & Co. KGaA;2010:[DOI](b) Huang K, Sun CL, Shi ZJ. Transition-metal-catalyzed C–C bond formation through the fixation of carbon dioxide. Chem Soc Rev. 2011;40(5):2435-2452.[DOI](c) He LN. Carbon dioxide chemistry. Beijing: Science Press; 2013(d) Centi G, Perathoner S. Green carbon dioxide: Advances in CO2 utilization. Hoboken: New York: John Wiley & Sons; 2014.[DOI](e) Maeda C, Miyazaki Y, Ema T. Recent progress in catalytic conversions of carbon dioxide. Catal Sci Technol. 2014;4(6):1482-1497.[DOI](f) Yu B, He LN. Upgrading carbon dioxide by incorporation into heterocycles. ChemSusChem. 2015;8(1):52-62.[DOI](g) Liu Q, Wu L, Jackstell R, Beller M. Using carbon dioxide as a building block in organic synthesis. Nat Commun. 2015;6:5933.[DOI](h) Yu D, Teong SP, Zhang Y. Transition metal complex catalyzed carboxylation reactions with CO2. Coord Chem Rev. 2015;293-294:279-291.[DOI](i) Song QW, Zhou ZH, He LN. Efficient, selective and sustainable catalysis of carbon dioxide. Green Chem. 2017;19(16):3707-3728.[DOI](j) Yuan G, Qi C, Wu W, Jiang H. Recent advances in organic synthesis with CO2 as C1 synthon. Curr Opin Green Sustain Chem. 2017;3:22-27.[DOI](k) Dabral S, Schaub T. The use of carbon dioxide (CO2) as a building block in organic synthesis from an industrial perspective. Adv Synth Catal. 2019;361(2):223-246.[DOI](l) Wang S, Xi C. Recent advances in nucleophile-triggered CO2-incorporated cyclization leading to heterocycles. Chem Soc Rev. 2019;48(1):382-404.[DOI](m) Velty A, Corma A. Advanced zeolite and ordered mesoporous silica-based catalysts for the conversion of CO2 to chemicals and fuels. Chem Soc Rev. 2023;52(5):1773-1946.[DOI]
-
3. (a) Vaitla J, Guttormsen Y, Mannisto JK, Nova A, Timo R, Bayer A, et al. Enantioselective incorporation of CO2: Status and potential. ACS Catal. 2017;7(10):7231-7244.[DOI](b) Ran CK, Chen XW, Gui YY, Liu J, Song L, Ren K, et al. Recent advances in asymmetric synthesis with CO2. Sci China Chem. 2020;63(10):1336-1351.[DOI](c) Shi Y, Pan BW, Zhou Y, Zhou J, Liu YL, Zhou F. Catalytic enantioselective synthesis using carbon dioxide as a C1 synthon. Org Biomol Chem. 2020;18(42):8597-8619.[DOI](d) Guo X, Wang Y, Chen J, Li G, Xia JB. Recent advances of CO2 fixation via asymmetric catalysis for the direct synthesis of optically active small molecules. Chin J Org Chem. 2020;40(8):2208-2220.[DOI]
-
4. (a) Maag H. Prodrugs of carboxylic acids. In: Stella VJ, Borchardt RT, Hageman MJ, Oliyai R, Maag H, Tilley JW, editors. Prodrugs. New York: Springer; 2007.[DOI](b) Lamberth C, Dinges J. Bioactive carboxylic compound classes: Pharmaceuticals and agrochemicals. Weinheim: Wiley-VCH Verlag GmbH & Co. KGaA; 2016.[DOI](c) Halama A, Zapadlo M. Synthesis, isolation, and analysis of stereoisomers of sacubitril. Org Process Res Dev. 2019;23(1):102-107.[DOI]
-
5. (a) Renslo AR, Luehr GW, Gordeev MF. Recent developments in the identification of novel oxazolidinone antibacterial agents. Bioorg Med Chem. 2006;14(12):4227-4240.[DOI](b) Zappia G, Menéndez P, Monache GD, Misiti D, Nevola L, Botta B. The contribution of oxazolidinone frame to the biological activity of pharmaceutical drugs and natural products. Mini Rev Med Chem. 2007;7(4):389-409.[DOI](c) Liu P, Jiang Y, Jiao L, Luo Y, Wang X, Yang T. Strategies for the discovery of oxazolidinone antibacterial agents: Development and future perspectives. J Med Chem. 2023;66(20):13860-13873.[DOI](d) Yuan S, Shen DD, Bai YR, Zhang M, Zhou T, Sun C, et al. Oxazolidinone: A promising scaffold for the development of antibacterial drugs. Eur J Med Chem. 2023;250:115239.[DOI]
-
6. (a) Alper H, Hamel N. Asymmetric synthesis of acids by the palladium-catalyzed hydrocarboxylation of olefins in the presence of (R)-(-)- or (S)-(+)-1,1’-binaphthyl-2,2’-diyl hydrogen phosphate. J Am Chem Soc. 1990;112(7):2803-2804.[DOI](b) Konrad TM, Fuentes JA, Slawin AMZ, Clarke ML. Highly enantioselective hydroxycarbonylation and alkoxycarbonylation of alkenes using dipalladium complexes as precatalysts. Angew Chem Int Ed. 2010;49(48):9197-9200.[DOI](c) Konrad TM, Durrani JT, Cobley CJ, Clarke ML. Simultaneous control of regioselectivity and enantioselectivity in the hydroxycarbonylation and methoxycarbonylation of vinyl arenes. Chem Commun. 2013;49(32):3306-3308.[DOI](d) Feng S, Buchwald SL. CuH-catalyzed regio- and enantioselective hydrocarboxylation of allenes: Toward carboxylic acids with acyclic quaternary centers. J Am Chem Soc. 2021;143(13):4935-4941.[DOI](e) Yao YH, Zou XJ, Wang Y, Yang HY, Ren ZH, Guan ZH. Palladium-catalyzed asymmetric Markovnikov hydroxycarbonylation and hydroalkoxycarbonylation of vinyl arenes: Synthesis of 2-arylpropanoic acids. Angew Chem Int Ed. 2021;60(43):23117-23122.[DOI]
-
7. (a) Stivala CE, Zakarian A. Highly enantioselective direct alkylation of arylacetic acids with chiral lithium amides as traceless auxiliaries. J Am Chem Soc. 2011;133(31):11936-11939.[DOI](b) Yu K, Miao B, Wang W, Zakarian A. Direct enantioselective and regioselective alkylation of β, γ-unsaturated carboxylic acids with chiral lithium amides as traceless auxiliaries. Org Lett. 2019;21(6):1930-1934.[DOI](c) Ju S, Kuzelka KP, Guo R, Krohn-Hansen B, Wu J, Nair SK, et al. A biocatalytic platform for asymmetric alkylation of α-keto acids by mining and engineering of methyltransferases. Nat Commun. 2023;14:5704.[DOI]
-
8. (a) Shevlin M, Friedfeld MR, Huaming S, Pierson NA, Hoyt JM, Campeau LC, et al. Nickel-catalyzed asymmetric alkene hydrogenation of α,β-unsaturated esters: High-throughput experimentation-enabled reaction discovery, optimization, and mechanistic elucidation. J Am Chem Soc. 2016;138(10):3562-3569.[DOI](b) Hoen R, Boogers JAF, Bernsmann H, Minnaard AJ, Meetsma A, Tiemersma-Wegman TD, et al. Achiral ligands dramatically enhance rate and enantioselectivity in the Rh/phosphoramidite-catalyzed hydrogenation of α, β-disubstituted unsaturated acids. Angew Chem Int Ed. 2005;44(27):4209-4212.[DOI](c) Zhu SF, Yu YB, Li S, Wang LX, Zhou QL. Enantioselective hydrogenation of α-substituted acrylic acids catalyzed by iridium complexes with chiral spiro aminophosphine ligands. Angew Chem Int Ed. 2012;51(35):8872-8875.[DOI](d) Hayashi T, Kawamura N, Ito Y. Asymmetric hydrogenation of trisubstituted acrylic acids catalyzed by a chiral (aminoalkyl)ferrocenylphosphine-rhodium complex. J Am Chem Soc. 1987;109(25):7876-7878.[DOI](e) Zhu SF, Zhou QL. Iridium-catalyzed asymmetric hydrogenation of unsaturated carboxylic acids. Acc Chem Res. 2017;50(4):988-1001.[DOI](f) Du X, Xiao Y, Huang JM, Zhang Y, Duan YN, Wang H, et al. Cobalt-catalyzed highly enantioselective hydrogenation of α,β-unsaturated carboxylic acids. Nat Commun. 2020;11(1):3239.[DOI](g) Zhong H, Shevlin M, Chirik PJ. Cobalt-catalyzed asymmetric hydrogenation of α,β-unsaturated carboxylic acids by homolytic H2 cleavage. J Am Chem Soc. 2020;142(11):5272-5281.[DOI](h) An C, Shaw MH, Tharp A, Verma D, Li H, Wang H, et al. Enantioselective enzymatic reduction of acrylic acids. Org Lett. 2020;22(21):8320-8325.[DOI]
-
9. (a) Ager DJ, Prakash I, Schaad DR. 1,2-amino alcohols and their heterocyclic derivatives as chiral auxiliaries in asymmetric synthesis. Chem Rev. 1996;96(2):835-876.[DOI](b) Gabriele B, Salerno G, Brindisi D, Costa M, Chiusoli GP. Synthesis of 2-oxazolidinones by direct palladium-catalyzed oxidative carbonylation of 2-amino-1-alkanols. Org Lett. 2000;2(5):625-627.[DOI](c) Overman LE, Remarchuk TP. Catalytic asymmetric intramolecular aminopalladation: enantioselective synthesis of vinyl-substituted 2-oxazolidinones, 2-imidazolidinones, and 2-pyrrolidinones. J Am Chem Soc. 2002;124(1):12-13.[DOI](d) Li P, Yuan X, Wang S, Lu S. A novel way to chiral 2-oxazolidinones: Selenium-catalyzed cyclocarbonylation of 2-aminoethanols. Tetrahedron. 2007;63(50):12419-12423.[DOI]
-
10. (a) Jung ME, Jung YH. Rapid synthesis of β-hydroxy-α-amino acids, such as L-threonine, β-hydroxyphenylalanine, and β-hydroxyleucine, via an application of the Sharpless asymmetric epoxidation. Tetrahedron Lett. 1989;30(48):6637-6640.[DOI](b) Iwama S, Katsumura S. A new route for protected amino alcohols from (R)-glycidol. Copper(I) mediated alkylation of 4-tosyloxymethyl-2-oxazolidinone. Bull Chem Soc Jpn. 1994;67(12):3363-3365.[DOI](c) Khan A, Xing J, Zhao J, Kan Y, Zhang W, Zhang YJ. Palladium-catalyzed enantioselective decarboxylative cycloaddition of vinylethylene carbonates with isocyanates. Chem Eur J. 2015;21(1):120-124.[DOI]
-
11. (a) Wang Q, Tan X, Zhu Z, Dong XQ, Zhang X. New synthetic strategy for chiral 2-oxazolidinones derivatives via rhodium-catalyzed asymmetric hydrogenation. Tetrahedron Lett. 2016;57(6):658-662.[DOI](b) Li W, Wollenburg M, Glorius F. Enantioselective synthesis of 2-oxazolidinones by ruthenium(ii)-NHC-catalysed asymmetric hydrogenation of 2-oxazolones. Chem Sci. 2018;9(29):6260-6263.[DOI](c) Liu Y, Yi Z, Yang X, Wang H, Yin C, Wang M, et al. Efficient access to chiral 2-oxazolidinones via Ni-catalyzed asymmetric hydrogenation: scope study, mechanistic explanation, and origin of enantioselectivity. ACS Catal. 2020;10(19):11153-11161.[DOI]
-
12. (a) Kolb HC, VanNieuwenhze MS, Sharpless KB. Catalytic asymmetric dihydroxylation. Chem Rev. 1994;94(8):2483-2547.[DOI](b) Kielland N, Whiteoak CJ, Kleij AW. Stereoselective synthesis with carbon dioxide. Adv Synth Catal. 2013;355(11-12):2115-2138.[DOI](c) Gómez JE, Kleij AW. Recent progress in stereoselective synthesis of cyclic organic carbonates and beyond. Curr Opin Green Sustain Chem. 2017;3:55-60.[DOI](d) Nie X, Ye CX, Ivlev SI, Meggers E. Nitrene-mediated C-H oxygenation: Catalytic enantioselective formation of five-membered cyclic organic carbonates. Angew Chem Int Ed. 2022;61(48):e202211971.[DOI]
-
13. Ghosh D, Dabas S, Kalose SA, Gotgi NM, Subramanian S. CO2 and asymmetry: Reviewing the advances in enantioselective catalytic strategies. Catal Sci Technol. 2025;15(3):618-646.[DOI]
-
14. Lu XB. Carbon Dioxide and Organometallics. In: Lu XB, editor. Topics in Organometallic Chemistry. Cham: Springer International Publishing; 2015. p. 171-198. Available from: https://link.springer.com/book/10.1007/978-3-319-22078-9
-
15. Childers MI, Longo JM, Van Zee NJ, LaPointe AM, Coates GW. Stereoselective epoxide polymerization and copolymerization. Chem Rev. 2014;114(16):8129-8152.[DOI]
-
16. Altava B, Cirujano FG, García-Verdugo E. Chiral catalysts for the enantioselective carbon dioxide-based cyclic carbonates and polycarbonates. Catalysts. 2023;13(11):1441.[DOI]
-
17. Suwankaisorn B, Kaewsai S, Natongchai W, D’ Elia V. Applications and synthesis of chiral cyclic carbonates from asymmetric CO2 conversion. Asian J Org Chem. 2025;14(7):e202500320.[DOI]
-
18. Guo Y, Wei L, Wen Z, Qi C, Jiang H. Recent progress on conversion of carbon dioxide into carbamates. Acta Phys Chim Sin. 2024;40(4):2307004.[DOI]
-
19. Chen XW, Yue JP, Wang K, Gui YY, Niu YN, Liu J, et al. Nickel-Catalyzed Asymmetric Reductive Carbo-Carboxylation of Alkenes with CO2. Angew Chem Int Ed. 2021;60(25):14068-14075.[DOI]
-
20. Cerveri A, Giovanelli R, Sella D, Pedrazzani R, Monari M, Nieto Faza O, et al. Enantioselective CO2 fixation via a heck-coupling/carboxylation cascade catalyzed by nickel. Chemistry. 2021;27(28):7657-7662.[DOI]
-
21. Zhang Q, Zhang H, Geng S, Zhao X, Liu S, Hong K, et al. Green transformation of CO2 into γ-amino alcohols with continuous stereocenters. Chem Commun. 2024;60(15):2062-2065.[DOI]
-
23. Zhang S, Li L, Li D, Zhou YY, Tang Y. Catalytic regio- and enantioselective boracarboxylation of arylalkenes with CO2 and diboron. J Am Chem Soc. 2024;146(5):2888-2894.[DOI]
-
24. Pettersen M, Do CD, Gorantla SM, Obst MF, Damm R, Putra AE, et al. Asymmetric boracarboxylation of styrenes using carbon dioxide. Adv Synth Catal. 2024;366(13):2976-2986.[DOI]
-
25. Zhou L, Li L, Zhang S, Kuang XK, Zhou YY, Tang Y. Catalytic regio- and enantioselective remote hydrocarboxylation of unactivated alkenes with CO2. J Am Chem Soc. 2024;146(28):18823-18830.[DOI]
-
26. Yu R, Huang L, Zhang H, Chen Y. Nickel-catalyzed hydrocarboxylation of styrenes with carbon dioxide and water. J Org Chem. 2025;90(26):9249-9252.[DOI]
-
27. Gui YY, Chen XW, Mo XY, Yue JP, Yuan R, Liu Y, et al. Cu-catalyzed asymmetric dicarboxylation of 1,3-Dienes with CO2. J Am Chem Soc. 2024;146(5):2919-2927.[DOI]
-
28. Wang MY, Jin X, Wang X, Xia S, Wang Y, Huang S, et al. Copper-catalyzed and proton-directed selective hydroxymethylation of alkynes with CO2. Angew Chem Int Ed. 2021;60(8):3984-3988.[DOI]
-
29. Li C, Chen XW, Liao LL, Gui YY, Yang JW, Zhang S, et al. Nickel-catalyzed atroposelective carbo-carboxylation of alkynes with CO2: En route to axially chiral carboxylic acids. Angew Chem Int Ed Engl. 2025;64(1):e202413305.[DOI]
-
30. Wang L, Li T, Perveen S, Zhang S, Wang X, Ouyang Y, et al. Nickel-catalyzed enantioconvergent carboxylation enabled by a chiral 2,2’-bipyridine ligand. Angew Chem Int Ed. 2022;61(51):e202213943.[DOI]
-
31. Zhou Q, Zhang C, Luo HL, Nie CX, Hu C, Ye JH, et al. Carbon-metal bond homolysis-recombination enabling enantioconvergent carboxylation with CO2: A theoretical study. Chin Chem Lett. 2026;37(5):111429.[DOI]
-
32. Hu Q, Wei B, Wang M, Liu M, Chen XW, Ran CK, et al. Enantioselective nickel-electrocatalyzed reductive propargylic carboxylation with CO2. J Am Chem Soc. 2024;146(21):14864-14874.[DOI]
-
33. Zhong Y, Jiang M, Yang D, Feng N, Sun Y, Wang H, et al. Nickel-catalyzed electrochemical carboxylation of propargylic esters with CO2 to 2,3-allenoic acids. Chin Chem Lett. 2025;36(12):111169.[DOI]
-
34. Chen XW, Li C, Gui YY, Yue JP, Zhou Q, Liao LL, et al. Atropisomeric carboxylic acids synthesis via nickel-catalyzed enantioconvergent carboxylation of aza-biaryl triflates with CO2. Angew Chem Int Ed. 2024;63(22):e202403401.[DOI]
-
35. Ye B, Su L, Zheng K, Gao S, Liu J. Synergistic photoredox/palladium catalyzed enantioconvergent carboxylation of racemic heterobiaryl (pseudo) halides with CO2. Angew Chem Int Ed. 2025;64(1):e202413949.[DOI]
-
36. Ye B, Lu Y, Yang X, Gao S, Liu J. Anion-suppression strategy enables palladium metallaphotoredox-catalyzed desymmetric carboxylation with CO2. J Am Chem Soc. 2026;148(11):12400-12410.[DOI]
-
37. Kang B, Wang L, Sun X, Liu H, Wen Z, Ren Y, et al. Enantioselective synthesis of axially chiral carbamates and amides with carbon dioxide via copper catalysis. Org Chem Front. 2023;10(20):5231-5241.[DOI]
-
38. Gao XT, Gan CC, Liu SY, Zhou F, Wu HH, Zhou J. Utilization of CO2 as a C1 building block in a tandem asymmetric A3 coupling-carboxylative cyclization sequence to 2-oxazolidinones. ACS Catal. 2017;7(12):8588-8593.[DOI]
-
39. Zhang Z, Zhang ZH, Zhou F, Zhou J. Catalytic enantioselective transfer hydrogenation-carboxylative cyclization to 4-fluoroalkyl 2-oxazolidinone with CO2 as the C1 synthon. Org. Lett., 2021;23(7):2726-2730.[DOI]
-
40. Gao XT, Xie SL, Zhou F, Wu HH, Zhou J. Multifunctional 1,3-diphenylguanidine for the carboxylative cyclization of homopropargyl amines with CO2 under ambient temperature and pressure. Chem Commun. 2019;55(95):14303-14306.[DOI]
-
41. Li ZH, Ma JS, Lu HY, Lin GQ, He ZT. Asymmetric multicomponent propargylations via carbon dioxide shuttling and fixation. ACS Catal. 2024;14(15):11646-11656.[DOI]
-
42. Lan Y, Liu P, Fang Z, Shao L, Cai Q, Wang X. Well-defined chiral dinuclear copper-catalyzed tandem asymmetric propargylic amination-carboxylative cyclization sequence toward chiral 2-oxazolidinone derivatives. Org Chem Front. 2024;11(22):6319-6326.[DOI]
-
43. Zhang Z, Sun Y, Gong Y, Tang DL, Luo H, Zhao ZP, et al. Enantioselective propargylic amination and related tandem sequences to α-tertiary ethynylamines and azacycles. Nat Chem. 2024;16(4):521-532.[DOI]
-
44. Zhang Z, Zhang ZH, Sun Y, Tang YH, Yang YZ, Zhou F, et al. Tandem asymmetric propargylic amination/carboxylative cyclization reaction to chiral 5-methylidene-2-oxazolidinones using CO2 as C1 synthon. Sci China Chem. 2025;68(4):1402-1411.[DOI]
-
45. Gong Y, Zhang Z, Liu H, Wang T, Jiang M, Feng N, et al. Trifluoroethanol-assisted asymmetric propargylic hydrazination to α-tertiary ethynylhydrazines enabled by sterically confined pyridinebisoxazolines. Nat Commun. 2025;16(1):4571.[DOI]
-
46. Nishiyori R, Mori T, Shirakawa S. Catalytic asymmetric CO2 utilization reaction for the enantioselective synthesis of chiral 2-oxazolidinones. Org Biomol Chem. 2023;21(19):4002-4006.[DOI]
-
47. Li X, Xue H, Yang S, Li Y, Dai C, Mai J, et al. Copper-catalyzed radical enantioconvergent C-O bond formation for asymmetric cross-coupling of allylamines and CO2. Angew Chem Int Ed. 2025;64(34):e202510560.[DOI]
-
48. Emelyanov MA, Stoletova NV, Smol’yakov AF, Il’in MM, Maleev VI, Larionov VA. Synthesis and a catalytic study of diastereomeric cationic chiral-at-cobalt complexes based on (R, R)-1,2-diphenylethylenediamine. Inorg Chem. 2021;60(18):13960-13967.[DOI]
-
49. Emelyanov MA, Lisov AA, Medvedev MG, Maleev VI, Larionov VA. Cobalt(III) complexes as bifunctional hydrogen-bond donor catalysts featuring halide anions for cyclic carbonate synthesis at ambient temperature and pressure: A mechanistic insight. Asian J Org Chem. 2022;11(5):e202100811.[DOI]
-
50. Emelyanov MA, Bachinskiy AV, Derkach YV, Chaliy VA, Smol’yakov AF, Medvedev MG, et al. Design of a chiral molecular pocket in a Ni(II) complex to improve stereoselectivity in the kinetic resolution of racemic epoxides with CO2. Tetrahedron Chem. 2024;12:100115.[DOI]
-
51. Esteve F, Porcar R, Bolte M, Altava B, Luis SV, García-Verdugo E. A bioinspired approach toward efficient supramolecular catalysts for CO2 conversion. Chem Catal. 2023;3(4):100482.[DOI]
-
52. Gao X, Zhong Y, Feng N, Sun Y, Yang D, Zhou F. Recent advances in electrochemical carboxylation of inert chemical bonds with carbon dioxide. Chin J Org Chem. 2024;44(10):3043-3062.[DOI]
-
53. Sun GQ, Liao LL, Ran CK, Ye JH, Yu DG. Recent advances in electrochemical carboxylation with CO2. Acc Chem Res. 2024;57(18):2728-2745.[DOI]
-
54. Wang Y, Yang G, Qiu Y. Organic electrochemical transformations of resource small molecules. Acc Chem Res. 2026;59(13):2117-2138.[DOI]
-
55. Mao B, Wei JS, Shi M. Recent advancements in visible-light-driven carboxylation with carbon dioxide. Chem Commun. 2022;58(67):9312-9327.[DOI]
-
56. Yan SS, Gao TY, Liu Y, Chen YF, Zuo JZ, Zhang QF, et al. Catalytic redox-neutral carboxylation with CO2. Chem Soc Rev. 2025;54(24):11583-11623.[DOI]
-
57. Luo H, Ren J, Sun Y, Liu Y, Zhou F, Shi G, et al. Recent advances in chemical fixation of CO2 based on flow chemistry. Chin Chem Lett. 2023;34(5):107782.[DOI]
-
58. Yan SS, Qi C, Gui Y, Li HR, Wang Y, Zhang W, et al. Recent advances in organic synthesis with CO2. Sci China Chem. 2026;69:4151-4222.[DOI]
Copyright
© The Author(s) 2027. This is an Open Access article licensed under a Creative Commons Attribution 4.0 International License (https://creativecommons.org/licenses/by/4.0/), which permits unrestricted use, sharing, adaptation, distribution and reproduction in any medium or format, for any purpose, even commercially, as long as you give appropriate credit to the original author(s) and the source, provide a link to the Creative Commons license, and indicate if changes were made.
Publisher’s Note
Science Exploration remains a neutral stance on jurisdictional claims in published
maps
and institutional affiliations. The views expressed in this article are solely those
of
the author(s) and do not reflect the opinions of the Editors or the publisher.
Share And Cite


Science Exploration Style
Yang D, Zhou H, Wang L, Wang H, Jiang M, Zhang Y, et al. Catalytic asymmetric synthesis of chiral carboxylic acids and heterocycles with CO2. Chiral Chem. 2027;3:202633. https://doi.org/10.70401/cc.2026.0038
Tips
Copy completed.
Submit a Manuscript
Author Instructions
Cite this Article
Article Metrics
0
View
0
Download
Cited
Article Updates

- Abstract
- Keywords
- 1. Introduction
- 2. Asymmetric Synthesis of Chiral Carboxylic Acids Involving CO2
- 3. Asymmetric Synthesis of Chiral Heterocycles Involving CO2
- 4. Summary and Outlook
- Authors contribution
- Conflicts of interest
- Ethical approval
- Consent to participate
- Consent for publication
- Availability of data and materials
- Funding
- References
- Copyright
 Get citation
Science Exploration Style

Yang D, Zhou H, Wang L, Wang H, Jiang M, Zhang Y, et al. Catalytic asymmetric synthesis of chiral carboxylic acids and heterocycles with CO2. Chiral Chem. 2027;3:202633. https://doi.org/10.70401/cc.2026.0038
copy
Share Link
copy